tm2619715-8_s1a - block - 29.994782s
As filed with the Securities and Exchange Commission on October 9, 2026.
Registration No. 333-299117
UNITED STATES
SECURITIES AND EXCHANGE COMMISSION
WASHINGTON, D.C. 20549
Amendment No. 1
to
FORM S-1
REGISTRATION STATEMENT
UNDER
THE SECURITIES ACT OF 1933
City Therapeutics, Inc.
(Exact name of registrant as specified in its charter)
| |
Delaware
(State or other jurisdiction of
incorporation or organization)
|
|
|
2834
(Primary Standard Industrial
Classification Code Number)
|
|
|
93-2762122
(I.R.S. Employer
Identification Number)
|
|
399 Binney Street
Cambridge, MA 02142
(857) 357-2024
(Address, including zip code, and telephone number, including area code, of registrant’s principal executive offices)
Andrew Orth, M.B.A.
President and Chief Executive Officer
City Therapeutics, Inc.
399 Binney Street
Cambridge, MA 02142
(857) 357-2024
(Name, address, including zip code, and telephone number, including area code, of agent for service)
Copies to:
| |
Kingsley L. Taft
Sarah Ashfaq
Goodwin Procter LLP
100 Northern Avenue
Boston, MA 02210
(617) 570-1000
|
|
|
Andrew Orth, M.B.A.
President and Chief Executive Officer
City Therapeutics, Inc.
399 Binney Street
Cambridge, MA 02142
(857) 357-2024
|
|
|
Darah Protas
Evan Leitner
Divakar Gupta
Richard Segal
Cooley LLP
1299 Pennsylvania Avenue, NW, Suite 700
Washington, DC 20004
(202) 842-7800
|
|
Approximate date of commencement of proposed sale to the public: As soon as practicable after this registration statement becomes effective.
If any of the securities being registered on this Form are to be offered on a delayed or continuous basis pursuant to Rule 415 under the Securities Act of 1933 check the following box: ☐
If this Form is filed to register additional securities for an offering pursuant to Rule 462(b) under the Securities Act, please check the following box and list the Securities Act registration statement number of the earlier effective registration statement for the same offering. ☐
If this Form is a post-effective amendment filed pursuant to Rule 462(c) under the Securities Act, check the following box and list the Securities Act registration statement number of the earlier effective registration statement for the same offering. ☐
If this Form is a post-effective amendment filed pursuant to Rule 462(d) under the Securities Act, check the following box and list the Securities Act registration statement number of the earlier effective registration statement for the same offering. ☐
Indicate by check mark whether the registrant is a large accelerated filer, an accelerated filer, a non-accelerated filer, a smaller reporting company or an emerging growth company. See the definitions of “large accelerated filer,” “accelerated filer,” “smaller reporting company,” and “emerging growth company” in Rule 12b-2 of the Exchange Act.
| |
Large accelerated filer
|
|
|
☐
|
|
|
Accelerated filer
|
|
|
☐
|
|
| |
Non-accelerated filer
|
|
|
☒
|
|
|
Smaller reporting company
|
|
|
☒
|
|
| |
|
|
|
|
|
|
Emerging growth company
|
|
|
☒
|
|
If an emerging growth company, indicate by check mark if the registrant has elected not to use the extended transition period for complying with any new or revised financial accounting standards provided pursuant to Section 7(a)(2)(B) of the Securities Act. ☐
The Registrant hereby amends this Registration Statement on such date or dates as may be necessary to delay its effective date until the Registrant shall file a further amendment which specifically states that this Registration Statement shall thereafter become effective in accordance with Section 8(a) of the Securities Act of 1933, as amended, or until the Registration Statement shall become effective on such date as the Securities and Exchange Commission, acting pursuant to said Section 8(a), may determine.
The information in this preliminary prospectus is not complete and may be changed. We may not sell these securities until the registration statement filed with the Securities and Exchange Commission is effective. This preliminary prospectus is not an offer to sell these securities and it is not soliciting an offer to buy these securities in any state or other jurisdiction where the offer or sale is not permitted.
Subject to Completion. Dated October 9, 2026.
9,722,222 Shares
Common Stock
This is an initial public offering of shares of common stock of City Therapeutics, Inc. We are offering 9,722,222 shares of our common stock.
Prior to this offering, there has been no public market for our common stock. It is currently estimated that the initial public offering price per share will be between $17.00 and $19.00. We have applied to list our common stock on the Nasdaq Global Market under the symbol “CTY.” We believe that upon the completion of this offering, we will meet the standards for listing on Nasdaq, and the completion of this offering is contingent upon such listing.
We are an “emerging growth company” and a “smaller reporting company” as defined under the U.S. federal securities laws and, as such, we have elected to comply with certain reduced reporting requirements in this prospectus. For additional information, see the section titled “Prospectus Summary — Implications of Being an Emerging Growth Company and a Smaller Reporting Company.”
See the section titled “Risk Factors” beginning on page 14 to read about factors that you should consider before deciding to invest in shares of our common stock.
Neither the Securities and Exchange Commission nor any state securities commission has approved or disapproved of these securities or determined if this prospectus is truthful or complete. Any representation to the contrary is a criminal offense.
| |
|
|
Per Share
|
|
|
Total
|
|
|
Initial public offering price
|
|
|
|
$ |
|
|
|
|
|
$ |
|
|
|
|
Underwriting discounts and commissions(1)
|
|
|
|
$ |
|
|
|
|
|
$ |
|
|
|
|
Proceeds, before expenses, to City Therapeutics, Inc.
|
|
|
|
$ |
|
|
|
|
|
$ |
|
|
|
(1)
See the section titled “Underwriting” for additional information regarding compensation payable to the underwriters.
We have granted the underwriters the option to purchase up to an additional 1,458,333 shares of common stock from us, at the initial public offering price, less the underwriting discounts and commissions.
The underwriters expect to deliver the shares against payment on or about , 2026.
| |
Goldman Sachs & Co. LLC
|
|
|
Jefferies
|
|
|
Stifel
|
|
| |
Oppenheimer & Co.
|
|
Prospectus dated , 2026
TABLE OF CONTENTS
| |
|
|
Page
|
|
|
|
|
|
|
|
1
|
|
|
|
|
|
|
|
|
10
|
|
|
|
|
|
|
|
|
12
|
|
|
|
|
|
|
|
|
14
|
|
|
|
|
|
|
|
|
95
|
|
|
|
|
|
|
|
|
97
|
|
|
|
|
|
|
|
|
99
|
|
|
|
|
|
|
|
|
100
|
|
|
|
|
|
|
|
|
102
|
|
|
|
|
|
|
|
|
105
|
|
|
|
|
|
|
|
|
126
|
|
|
|
|
|
|
|
|
186
|
|
|
|
|
|
|
|
|
197
|
|
|
|
|
|
|
|
|
209
|
|
|
|
|
|
|
|
|
212
|
|
|
|
|
|
|
|
|
216
|
|
|
|
|
|
|
|
|
219
|
|
|
|
|
|
|
|
|
225
|
|
|
|
|
|
|
|
|
227
|
|
|
|
|
|
|
|
|
231
|
|
|
|
|
|
|
|
|
239
|
|
|
|
|
|
|
|
|
239
|
|
|
|
|
|
|
|
|
239
|
|
|
|
|
|
|
|
|
F-1
|
|
|
Neither we nor the underwriters have authorized anyone to provide you with any information or make any representations other than those contained in this prospectus or in any free writing prospectuses prepared by or on behalf of us or to which we have referred you. We and the underwriters take no responsibility for, and can provide no assurance as to the reliability of, any other information that others may give you. We and the underwriters are not making an offer to sell these securities in any jurisdiction where the offer or sale is not permitted. You should assume that the information appearing in this prospectus or in any applicable free writing prospectus is current only as of its date, regardless of its time of delivery or any sale of shares of our common stock. Our business, financial condition, results of operations and prospects may have changed since that date.
For investors outside of the U.S., we have not, and the underwriters have not, done anything that would permit this offering or possession or distribution of this prospectus in any jurisdiction where action for that purpose is required, other than the U.S. Persons outside of the U.S. who come into possession of this prospectus must inform themselves about, and observe any restrictions relating to, the offering of the shares of our common stock and the distribution of this prospectus outside of the U.S.
We own, have applied for or have rights to use one or more registered and common law trademarks, service marks and/or trade names in connection with our business in the U.S., which may be used throughout this prospectus. This prospectus also includes trademarks, trade names, and service marks of third parties which are the property of their respective owners. Our use or display of third parties’ trademarks, service marks, trade names, or products in this prospectus is not intended to, and does not
imply a relationship with, or endorsement or sponsorship by us. Solely for convenience, the trademarks, service marks, logos and trade names referred to in this prospectus may appear without the ®, TM or SM symbols, but the omission of such references is not intended to indicate, in any way, that we will not assert, to the fullest extent under applicable law, our rights, or the rights of the applicable owner of or licensor to these trademarks, service marks, and trade names.
Market, Industry and Other Data
The market data and certain other statistical information used throughout this prospectus are based on independent industry publications, governmental publications, reports by market research firms or other independent sources that we believe to be reliable sources. Industry publications and third-party research, surveys and studies generally indicate that their information has been obtained from sources believed to be reliable, although they do not guarantee the accuracy or completeness of such information. We are responsible for all of the disclosure contained in this prospectus, and we believe that these sources are reliable; however, we have not independently verified the information contained in such publications. While we are not aware of any misstatements regarding any third-party information presented in this prospectus, their estimates, in particular, as they relate to projections, involve numerous assumptions, are subject to risks and uncertainties, and are subject to change based on various factors, including those discussed under the section titled “Risk Factors” and elsewhere in this prospectus. Some data are also based on our good faith estimates.
PROSPECTUS SUMMARY
This summary highlights selected information contained elsewhere in this prospectus. This summary does not contain all of the information you should consider before investing in our common stock. You should read this entire prospectus carefully, including the sections of this prospectus titled “Risk Factors,” “Cautionary Note Regarding Forward-Looking Statements,” “Management’s Discussion and Analysis of Financial Condition and Results of Operations” and our consolidated financial statements and the related notes included elsewhere in this prospectus, before making an investment decision.
Unless otherwise indicated, all references in this prospectus to “City,” “City Therapeutics,” the “company,” “we,” “our,” “us” or similar terms refer to City Therapeutics, Inc. and its wholly owned subsidiary, or either or both as the context may require. References in this prospectus to program and financial guidance as occurring in “early,” “mid-” or “late,” as applicable, in a given year refer to the first and second quarters, the second and third quarters, and the third and fourth quarters, respectively, of the applicable calendar year.
Overview
We are a clinical-stage biotechnology company developing the next generation of RNA interference (“RNAi”) therapeutics designed to silence the expression of disease-relevant proteins, with the aim of delivering meaningful and durable benefits for patients in need. We believe our deep RNAi expertise and City Platform capabilities uniquely position us to achieve our goal of developing potentially best-in-class therapeutics. Our vision is to lead the next generation of RNAi therapeutics, and our mission is to improve patients’ lives through breakthrough RNAi science as we build a biopharmaceutical company with enduring impact.
Our current pipeline is focused on indications with significant unmet medical need, including thromboembolic diseases, Stargardt disease type 1 (“Stargardt disease”), and anemia of chronic disease, where we believe disease biology supports RNAi-mediated knockdown as a differentiated approach compared to other modalities. Led by pioneers and experts in the field of RNAi, we have made rapid progress since founding our company in July 2023, discovering and advancing three programs into or toward the clinic in under three years, and our goal is for our City Platform to deliver one to two new investigational new drug application (“IND”) or IND-equivalent applications per year.
Several members of our leadership team, including John Maraganore, Ph.D., our co-founder and Executive Chair and former founding Chief Executive Officer of Alnylam Pharmaceuticals, Inc. (“Alnylam”), and Tracy Zimmermann, Ph.D., our Chief Scientific Officer, played a foundational role in establishing RNAi therapeutics as a recognized treatment modality, as evidenced by multiple FDA-approved RNAi therapeutics that each of them worked in development on, and our President and Chief Executive Officer, Andrew (Andy) Orth, M.B.A., led the launch of multiple RNAi therapeutics, including the first U.S. Food and Drug Administration (“FDA”)-approved RNAi therapeutic. Collectively, the team at City has contributed to the discovery, development, or commercialization of seven of the eight currently FDA-approved RNAi therapeutics, and 75% of our research leaders have prior RNAi therapeutics discovery or development experience.
RNAi is a naturally occurring biological process that regulates or silences gene expression by “interfering” with messenger RNA (“mRNA”) before it can be translated into protein. RNAi has inherent advantages as a therapeutic mechanism because it is a highly specific, potent, and durable approach to selectively silence the expression of disease-relevant proteins, including intracellular targets that are inaccessible to traditional small molecules and antibodies. In addition, RNAi therapeutics benefit from a simpler molecular architecture relative to biologics, consisting of short, chemically synthesized oligonucleotides rather than large, complex proteins, which enables more consistent and scalable manufacturing. RNAi has been established mechanistically and commercially, with the first of the eight FDA-approved RNAi therapeutics coming to market in 2018 and multiple products achieving substantial and growing commercial sales. With approximately 100 programs currently in clinical development, we believe RNAi therapeutics are poised to follow a growth trajectory analogous to that of monoclonal
antibody therapeutics, a modality that was first approved by the FDA in the mid-1990s and has since expanded to more than 150 approved medicines.
While RNAi is an established therapeutic modality, we are early in our development efforts, and our City Platform and approach to the engineering and development of certain of our product candidates is unproven and incorporates novel technologies that may not result in marketable products. Our lead product candidate, CITY-FXI, is in an early stage of clinical development, and our other product candidates and development programs are in preclinical development or drug discovery. We have only recently initiated our first clinical trial, have not yet completed any clinical trials, and have limited experience conducting and managing clinical trials. None of our product candidates has been approved for commercial sale, we have not generated revenue from product sales, and we have not yet demonstrated an ability to successfully complete clinical development or obtain marketing approval for and commercialize any of our product candidates. Our product candidates may fail in development or never receive regulatory approval, and we will require substantial additional capital to complete development of and, if approved, commercialize our product candidates. See the sections titled “—Summary of Material Risks Associated With Our Business” and “Risk Factors” for a discussion of these and other risks and uncertainties associated with our business.
Our Pipeline
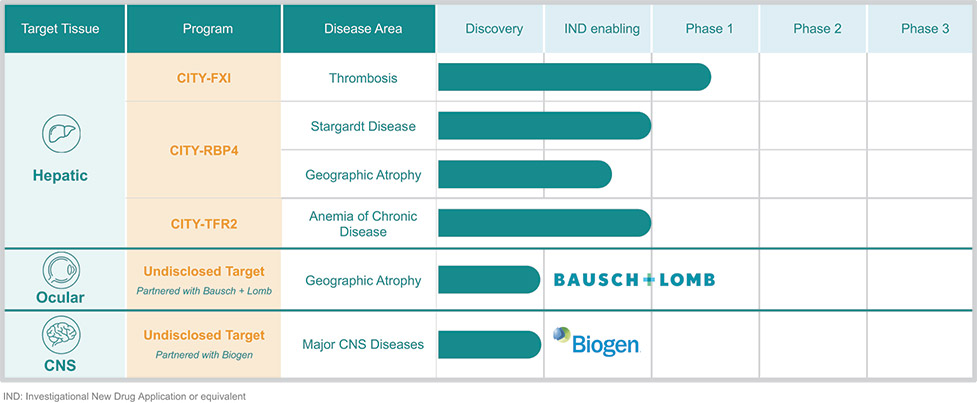
We are advancing a pipeline of RNAi therapeutics designed to achieve potent, specific, and durable suppression of disease-relevant hepatic and extrahepatic targets to treat diseases with significant unmet medical needs. The following table summarizes our current development stage programs:
All of our product candidates are in early clinical or preclinical development. We have not demonstrated that any of our product candidates are safe or effective or provide advantages over existing or investigational therapies, and results of preclinical studies, including our non-human primate (“NHP”) studies, may not be predictive of results in clinical trials. None of our product candidates has been approved by the FDA or any comparable foreign regulatory authority, and we may never obtain regulatory approval for any of our product candidates.
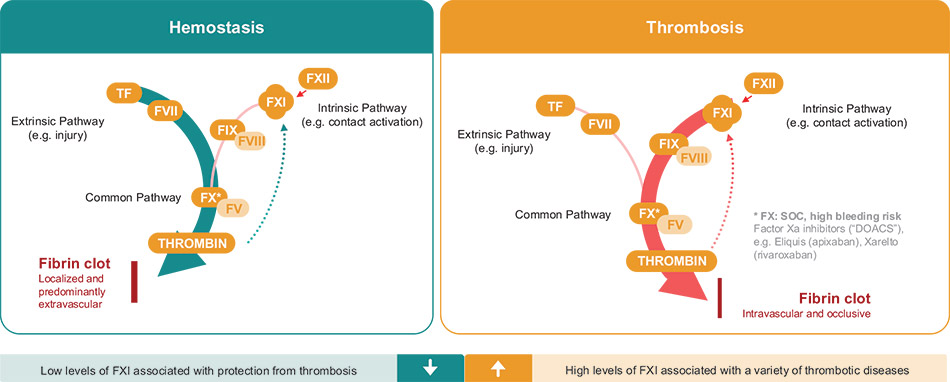
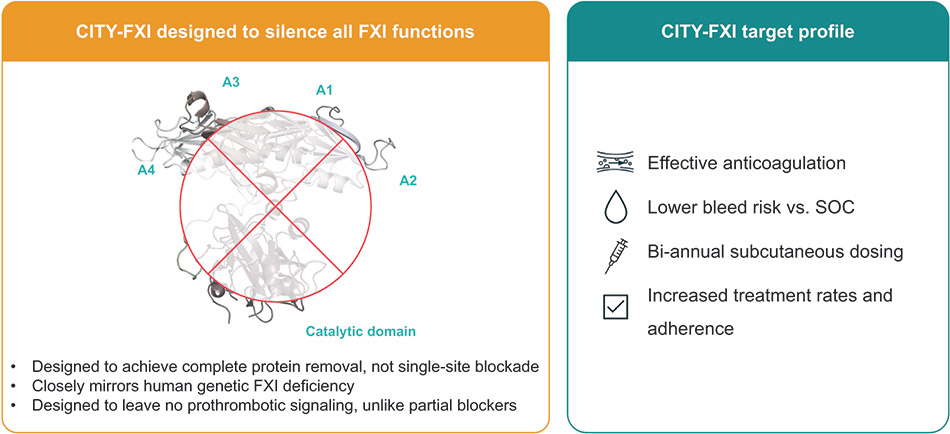
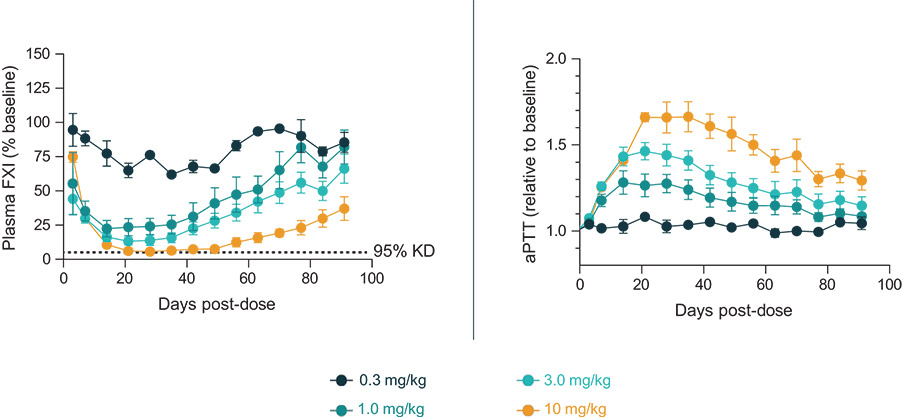
Our most advanced product candidate, CITY-FXI, is a City-engineered N-acetylgalactosamine (“GalNAc”)-conjugated small interfering RNA (“siRNA”) therapeutic targeting Factor XI (“FXI”). CITY-FXI illustrates our strategy of engineering highly potent and specific RNAi triggers directed at therapeutic targets supported by third-party human genetic and clinical data. We are advancing CITY-FXI for the prevention of thromboembolic diseases based on its potential to reduce FXI activity and thereby prevent thromboembolic events while mitigating bleeding risk. Thrombosis, the formation of harmful clots inside a blood vessel, is responsible for approximately one in four deaths worldwide. FXI is a genetically and clinically supported target that contributes to pathological thrombosis while playing a limited role in hemostasis, or normal blood clotting. In preclinical studies with CITY-FXI, we observed 95% FXI
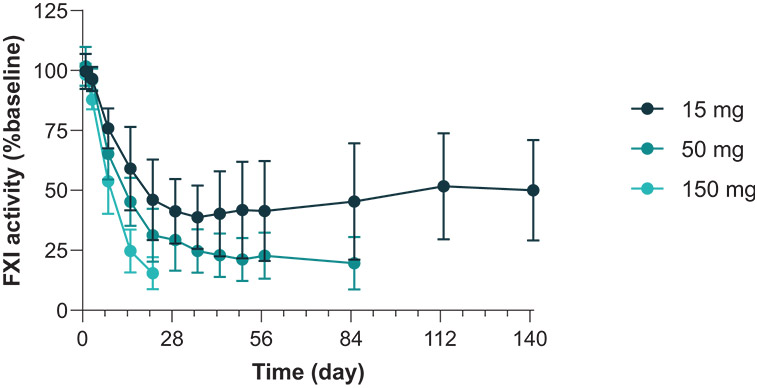
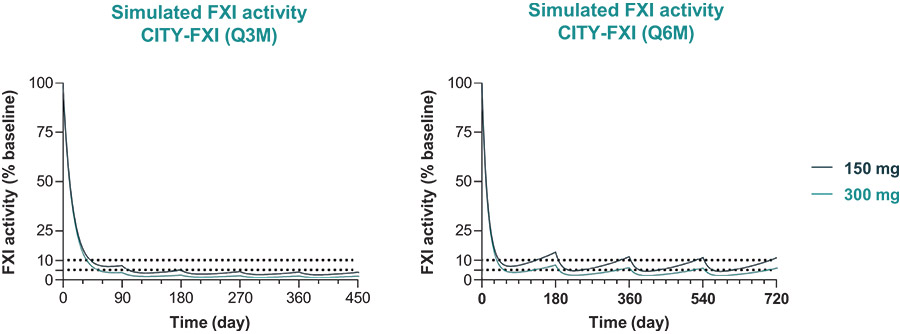
knockdown with prolonged clotting time in NHPs and no observed off-target effects in cell-based assays. We initiated our Phase 1 clinical trial for CITY-FXI in 2026, and initial data from the first three cohorts of the single-ascending dose (“SAD”) portion of our study in healthy volunteers showed that CITY-FXI was generally well-tolerated by participants with greater than 60% FXI knockdown observed following a single 15 mg subcutaneous dose, close to 80% FXI knockdown after a single 50 mg subcutaneous dose, and 85% FXI knockdown three weeks after a single 150 mg subcutaneous dose. We expect to report additional data from our Phase 1 study in late 2026. We intend to develop CITY-FXI for thromboembolic disease indications, including secondary stroke prevention, direct oral anticoagulant (“DOAC”)-ineligible atrial fibrillation (“AFib”), extended secondary prophylaxis following venous thromboembolism (“VTE”), and cancer-associated thrombosis, specifically in patients for whom DOACs are often not used because of heightened bleeding risk. We plan to first evaluate efficacy and safety in thrombotic disease through a Phase 2 trial in total knee arthroplasty (“TKA”), a clinical setting in which patients are at increased risk of postoperative thrombosis. We expect to initiate the Phase 2 study in TKA in mid-2027.
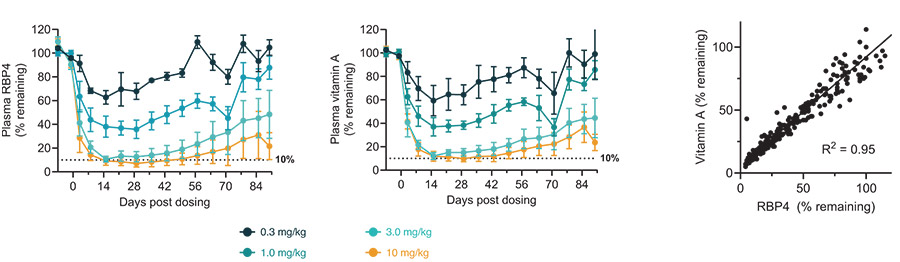
Our second product candidate, CITY-RBP4, is a City-engineered GalNAc-conjugated siRNA therapeutic, in development for the treatment of Stargardt disease, which is the most common inherited macular degenerative disease in children and young adults. There are currently no approved therapies for the disease. More than 30,000 people in the United States are diagnosed with Stargardt disease, and some studies suggest that the prevalence is even higher, indicating there could be a significant number of undiagnosed patients who emerge upon FDA approval of a treatment. Stargardt disease is debilitating, and many patients become legally blind in their late twenties. Patients face decades of progressive loss of central vision that renders daily activities, including reading, recognizing faces, and driving impossible. Stargardt disease is caused by mutations in the ABCA4 gene, which lead to the toxic accumulation of vitamin A byproducts in the eye. CITY-RBP4 targets retinol binding protein 4 (“RBP4”), the primary carrier of retinol, a form of vitamin A, that transports retinol from the liver to the eye. Inhibition of RBP4 has been supported by third-party clinical proof-of-mechanism data for the treatment of Stargardt disease. In addition, in a preclinical study in NHPs, following treatment with CITY-RBP4, we observed approximately 90% suppression of circulating vitamin A and RBP4 at dose levels of 3 mg/kg and 10 mg/kg. In mid-2026, we filed an application for CITY-RBP4 to initiate a Phase 1 clinical trial in late 2026 and we expect to report initial data from this trial in mid-2027. We are also exploring CITY-RBP4 in adjacent indications, including Geographic Atrophy (“GA”) secondary to age-related macular degeneration. We expect to initiate a Phase 2 clinical trial for CITY-RBP4 in GA in late 2027.
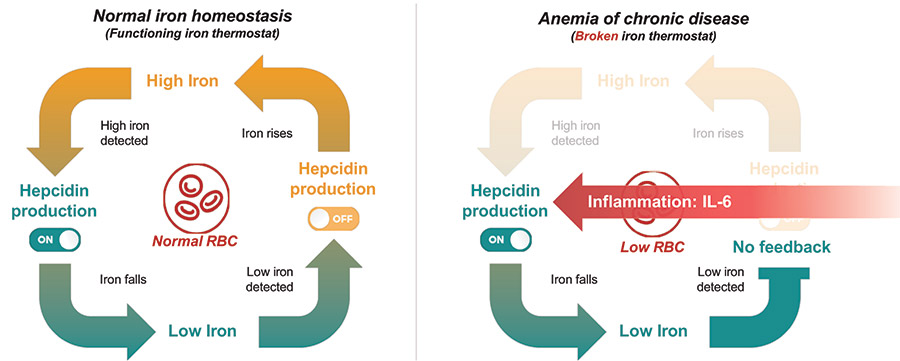
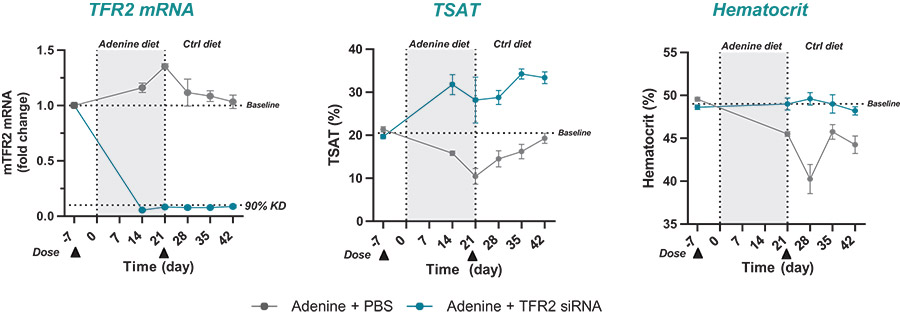
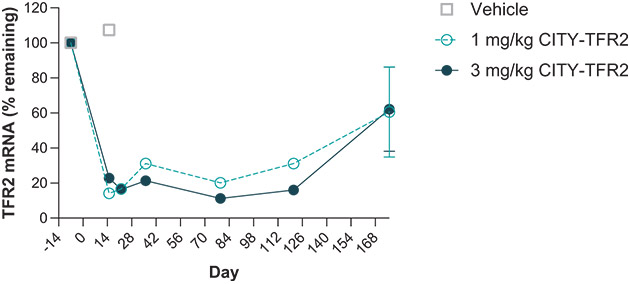
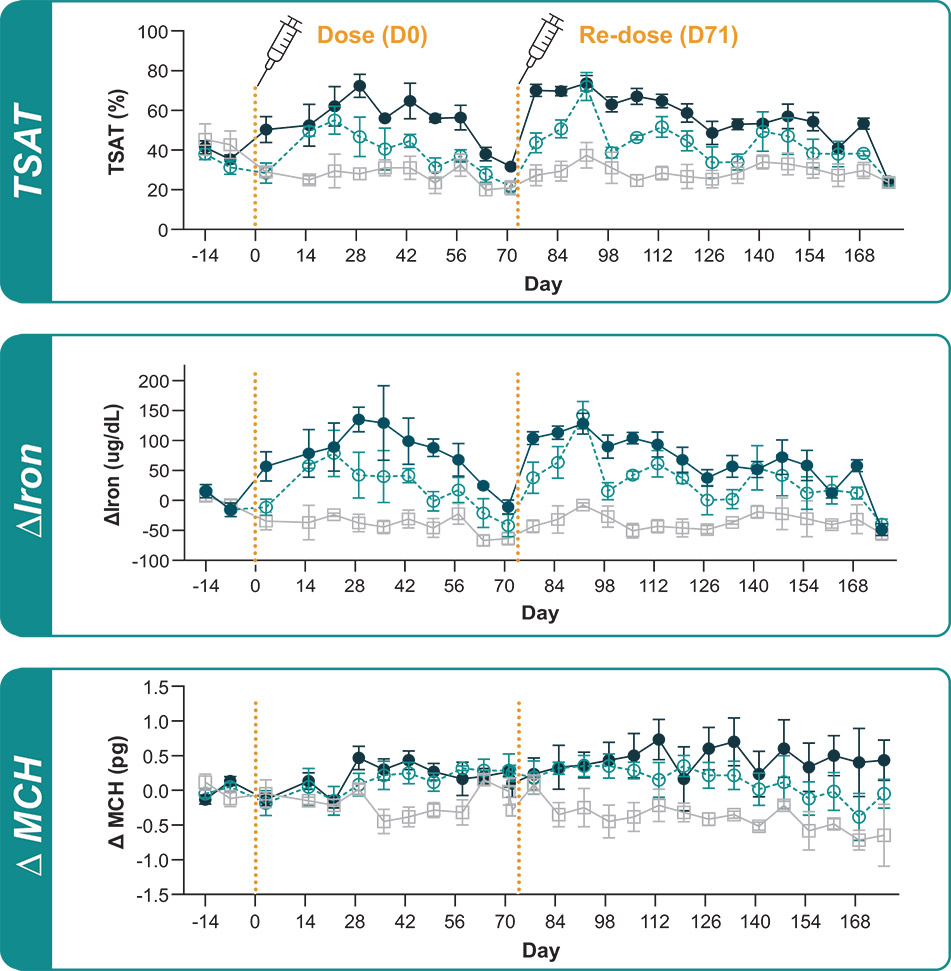
Our third program, CITY-TFR2, is a City-engineered GalNAc-conjugated siRNA therapeutic candidate in development for anemia of chronic disease, which accounts for up to 40% of all anemias and affects more than five million people in the United States. In the inflammatory states associated with anemia of chronic disease, the body increases production of hepcidin, the hormone controlling systemic iron, which restricts iron availability and starves red blood cell production. CITY-TFR2 is designed to silence transferrin receptor 2 (“TFR2”), a protein predominantly produced in the liver that is critical to the regulation of hepcidin production. This lowers hepcidin, restoring the iron supply needed to correct anemia. In a preclinical study in NHPs, following treatment with CITY-TFR2, we observed approximately 90% TFR2 mRNA knockdown with a concomitant increase in serum iron. We are initially focused on development of CITY-TFR2 in anemia associated with myelofibrosis, as anemia is a major contributor to the clinical burden of the disease. An investigator-initiated trial (“IIT”) for CITY-TFR2 is expected to begin in China in late 2026 and we expect to report data from the IIT in mid-2027. We also plan to file an IND or IND-equivalent application in late 2026 and initiate a Phase 1/2 study for CITY-TFR2 in early 2027, with potential to expand into other chronic conditions where anemia is prevalent, such as chronic kidney disease and inflammatory diseases, including inflammatory bowel disease.
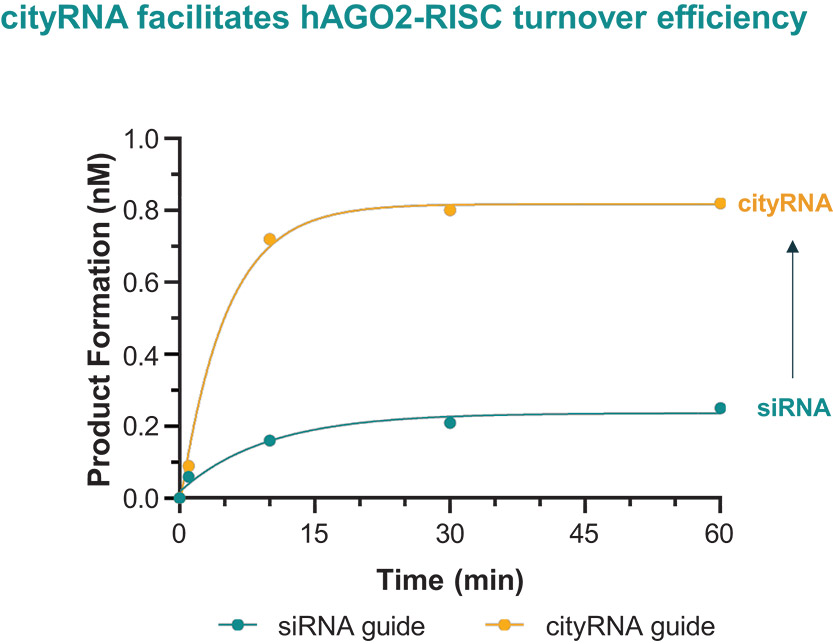
Beyond our lead programs, we are advancing discovery candidates designed to expand the reach of RNAi therapeutics across new tissues, targets and disease areas. These candidates include a fourth liver-directed program, CITY-L04, directed at an undisclosed target, as well as two discovery-stage programs based on our proprietary cleavage-inducing tiny RNAs (“cityRNAs”), CITY-O02 and CITY-C02, which are focused on ocular and CNS indications, respectively.
We have formed strategic collaborations with Bausch + Lomb Ireland Limited (“Bausch + Lomb”) and Biogen MA Inc. (“Biogen”) to advance extrahepatic RNAi therapeutics for diseases of the eye and the central nervous system (“CNS”), respectively. We believe these collaborations reinforce the potential of and enhance our City Platform, particularly our next-generation RNAi trigger and delivery technologies, while preserving our freedom to pursue additional targets in the eye and CNS and our rights to any City Platform innovations developed under the collaborations. In July 2024, we entered into a research collaboration and license agreement with Bausch + Lomb (the “B+L Agreement”) for a single target, focused on developing a novel RNAi therapy for retinal diseases, including GA. The collaboration leverages our next-generation delivery technologies to enable intravitreal delivery of a novel RNAi therapeutic, and we retain rights to the delivery technology we build under the agreement. We plan to substantially complete the research and development under the B+L Agreement to enable Bausch + Lomb to select a development candidate by late 2026. In May 2025, we entered into a research collaboration and license agreement with Biogen (the “Biogen Agreement”) focused on the discovery and development of an RNAi program for neurodegenerative diseases. The collaboration leverages our trigger engineering expertise and Biogen’s proprietary drug delivery technology to develop a systemic RNAi therapeutic for certain CNS diseases. The collaboration will initially focus on a single target, and Biogen has the option to designate one additional target for a separate collaboration and license agreement on substantially the same terms. We plan to substantially complete the research and development under the Biogen Agreement by late 2026 to enable Biogen to select a development candidate by late 2027.
Our City Platform
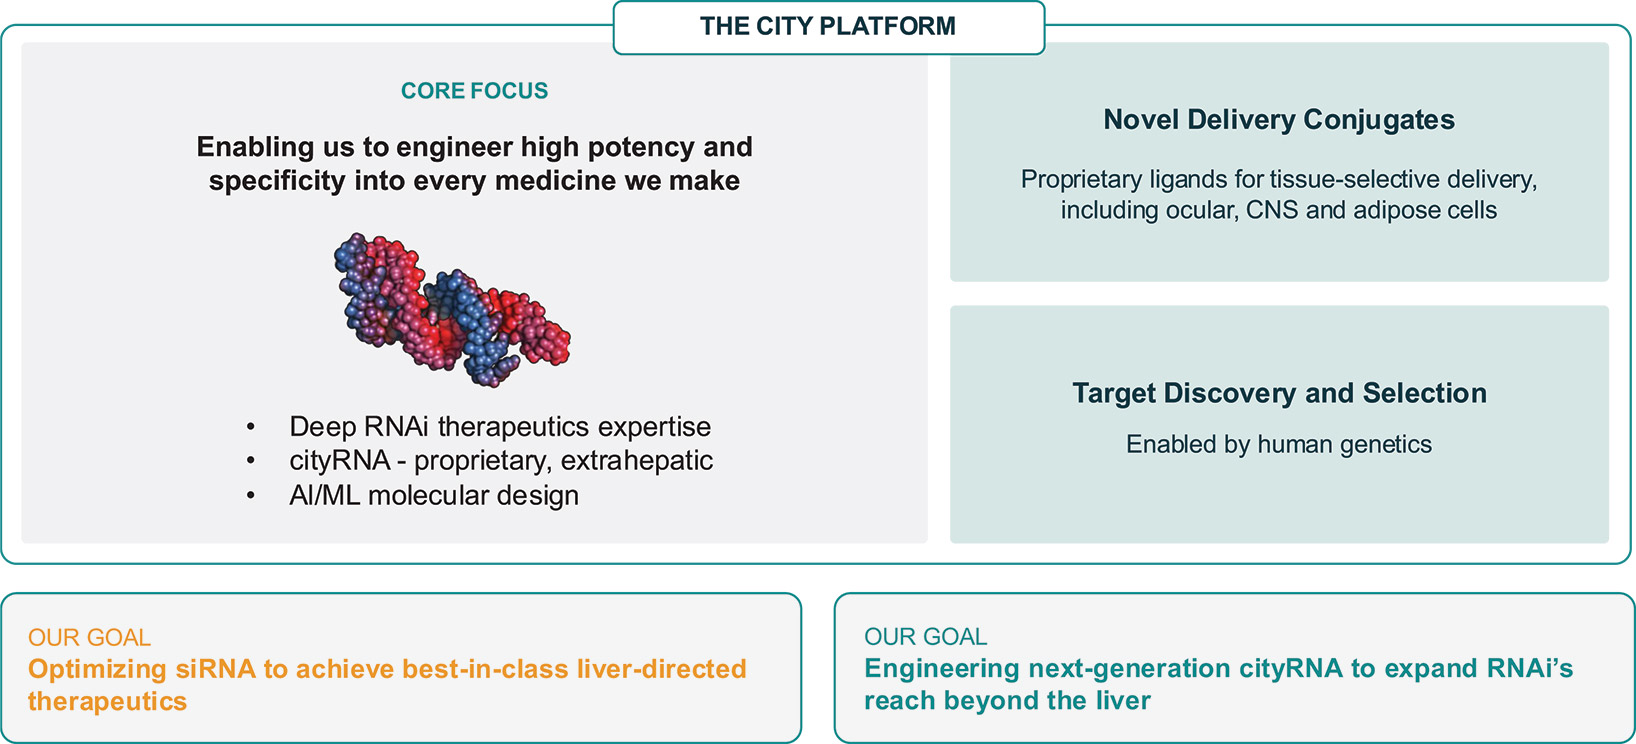
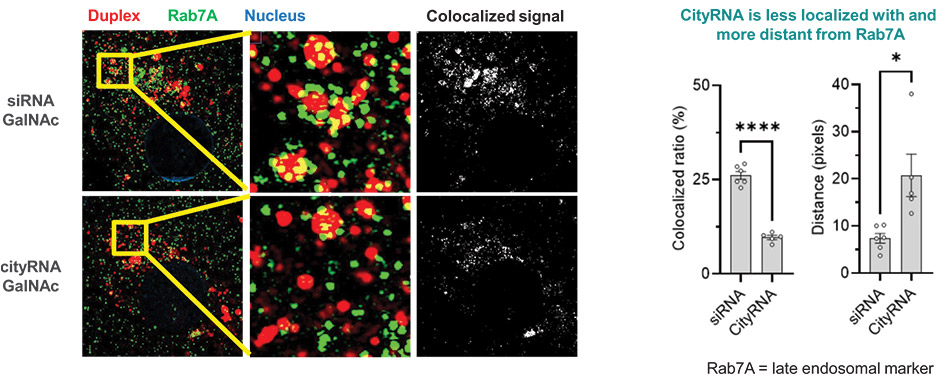
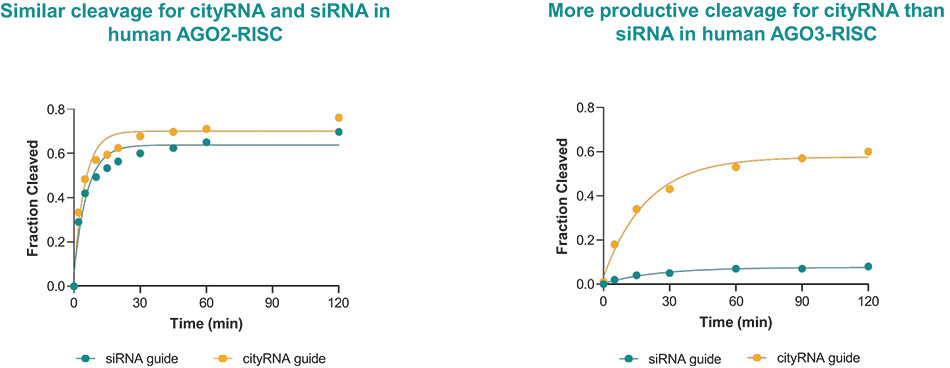
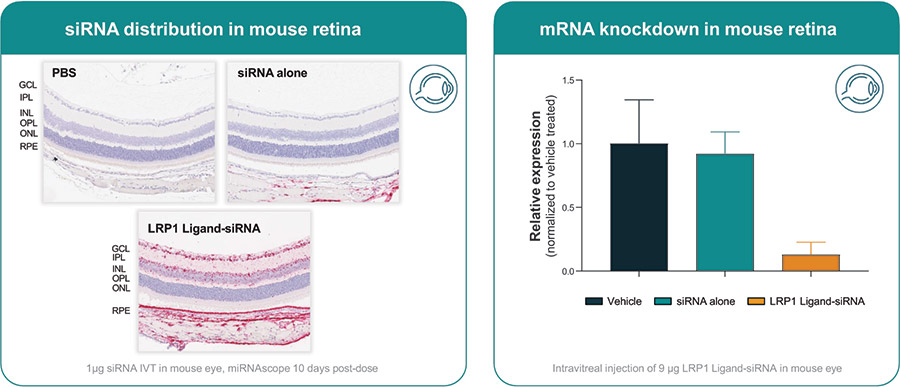
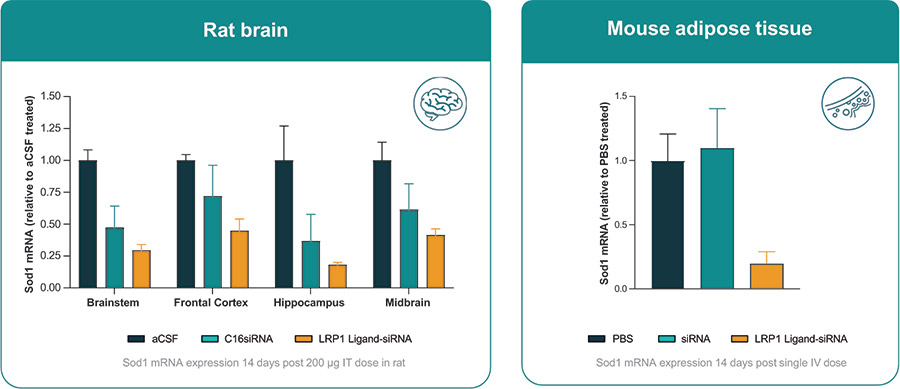
Our City Platform is the engine behind every therapeutic we develop. Our City Platform is designed to integrate three core capabilities: RNAi trigger engineering, novel delivery conjugates designed to enable tissue selectivity, and target discovery and selection enabled by our insights into human genetics. Critical to every RNAi medicine is the trigger that harnesses the cellular RNAi machinery to silence a disease-relevant gene. We believe our expertise enables us to engineer the trigger for high potency and specificity, and our novel cityRNA triggers are a differentiated approach to unlock extrahepatic delivery. Our proprietary artificial intelligence and machine learning (“AI/ML”)-enabled trigger design capability uses a large RNAi activity dataset to predict high-potential trigger molecules in silico, accelerating design cycles from months to weeks. We conjugate our triggers with tissue-targeting ligands for precise delivery to cell types in and beyond the liver, starting with our proprietary low-density lipoprotein receptor-related protein 1 (“LRP1”) ligands, which are designed to enable delivery to ocular, CNS, and adipose tissues. Our pipeline programs are selected through a genetics-led target discovery and selection capability that leverages insights from human biology and other sources of substantiation, including clinical data. We use this capability to identify differentiated and genetically supported targets where RNAi is designed to confer a durable therapeutic advantage. By uniting these capabilities in a single platform, our goal is to systematically create and develop potentially best-in-class therapeutics, with plans to file one to two new IND or IND-equivalent applications per year. The filing of an IND or IND-equivalent application, or the clearance of any such application by the FDA or a comparable foreign regulatory authority, is not indicative of the future success of any of our product candidates and does not increase the likelihood that any of our product candidates will receive regulatory approval. Given our early stage of development, it will take many years before we complete clinical development of and receive regulatory approval for any of our product candidates, if ever.
Certain foundational elements of our City Platform are supported by intellectual property licensed from the Ohio State Innovation Foundation (“OSIF”). In October 2023, we entered into a license agreement (as amended, the “OSIF License Agreement”) with OSIF, pursuant to which OSIF granted us an exclusive royalty-bearing license to certain patent rights covering the originating technology underlying cityRNAs and certain other aspects of our technology platform.
Our Strategy
We are driven by the opportunity to bring durable, meaningful benefit to patients through the next generation of RNAi therapeutics. Anchored by deep scientific expertise and our proprietary City Platform, our goal is to develop potentially best-in-class therapeutics and to build a lasting, fully integrated RNAi
company capable of delivering transformative therapeutics across a multitude of diseases. We intend to pursue this goal through the following key strategies:
•
Pioneer the future of RNAi trigger design and delivery.
•
Advance development candidates in high unmet medical need indications where RNAi offers meaningful advantages.
•
Selectively pursue value-accretive partnerships for certain programs to maximize the value of our portfolio.
•
Bring exceptional depth of experience to accelerate and expand our platform and pipeline.
Our Team and History
Defining the next generation of RNAi therapeutics requires a unique combination of scientific, clinical, and commercial expertise. We have assembled a team with deep experience across each of these disciplines. Collectively, our team has contributed to the discovery, development, or commercialization of seven of the eight currently FDA-approved RNAi therapeutics. Our Executive Chair and co-founder, John Maraganore, Ph.D., is a pioneer in the field of RNAi therapeutics who served as the former founding Chief Executive Officer of Alnylam for nearly 20 years, where he was instrumental in advancing the company from early platform research through the development, global regulatory approval, and commercialization of the first five RNAi therapeutics, and previously held senior leadership roles at Millennium Pharmaceuticals, Inc. and Biogen Inc. Our President and Chief Executive Officer, Andrew (Andy) Orth, M.B.A., brings more than 25 years of industry experience and held multiple leadership roles at Alnylam, including supporting the launch of ONPATTRO, the first FDA-approved RNAi therapeutic, before serving as Chief Commercial Officer of Krystal Biotech, Inc. Our Chief Medical Officer, Baisong Mei, M.D., Ph.D., a physician scientist and drug developer with 25 years of experience in the pharmaceutical and biotech industry, played a critical leadership role in the development and regulatory approval of ALPROLIX, ELOCTATE, ALTUVIIIO, and the siRNA medicine, QFITLIA. Our Chief Scientific Officer, Tracy Zimmermann, Ph.D., was the lead author of a landmark 2006 study, published in Nature, which provided the first-ever demonstration of RNAi-mediated gene silencing in NHPs and contributed heavily to the development of several approved RNAi therapeutics, including as co-inventor of AMVUTTRA.
Our board of directors consists of highly experienced biotechnology executives and investors, including our Executive Chair, John Maraganore, Ph.D., former founding Chief Executive Officer and founding member of the board of directors of Alnylam, Barry Greene, former President and Chief Executive Officer of Sage Therapeutics, Inc. and former long-time Chief Operating Officer of Alnylam, Robert Nelsen, M.B.A., co-founder and Managing Director of ARCH Venture Partners, Saraswathy (Sara) V. Nochur, Ph.D., former Chief Regulatory Officer and Chief Diversity, Equity and Inclusion Officer of Alnylam, Ron Philip, former Chief Executive Officer and a member of the board of directors of Orbital Therapeutics, Inc., and Andy Orth, our President and Chief Executive Officer. Further, we have assembled a cross-functional scientific and drug discovery advisory board, comprised of seasoned drug developers and leading academic scientists at the forefront of RNAi therapeutics.
From our inception in July 2023 through June 30, 2026, we have raised approximately $238.8 million in equity capital from a syndicate of premier life sciences investors, including ARCH Venture Partners, Fidelity Management & Research Company, Invus, and Viking Global Investors. Prospective investors should not rely on the investment decisions of our existing investors, as these investors may have different risk tolerances and, in certain cases, have received their shares in private financings at prices lower than the price offered to the public in this offering. See the sections titled “Certain Relationships and Related Person Transactions” and “Principal Stockholders” for more information on prior purchases by and current holdings of certain of these investors.
Summary of Material Risks Associated With Our Business
Our business is subject to a number of risks of which you should be aware before making an investment decision. These risks include, but are not limited to, the following:
•
We are a clinical-stage biotechnology company with a limited operating history and no products approved by regulators for commercial sale, which may make it difficult to evaluate our current and future business prospects.
•
The approach we are taking to discover and develop certain of our product candidates is novel and may never lead to marketable products.
•
We are early in our development efforts. We have only recently initiated early clinical studies, and as a result it will be years before we commercialize a product candidate, if ever. If we are unable to identify and advance product candidates through preclinical studies and clinical trials, obtain marketing approval and ultimately commercialize them, or experience significant delays in doing so, our business will be materially harmed.
•
We have incurred significant net losses since our inception, including net losses of $27.8 million and $58.5 million for the years ended December 31, 2024 and 2025, respectively, and $24.3 million and $53.4 million for the six months ended June 30, 2025 and 2026, respectively. As of June 30, 2026 we had an accumulated deficit of $142.5 million. We anticipate that we will continue to incur significant losses for the foreseeable future.
•
Even if this offering is successful, we will require substantial additional capital to finance our operations in the future. If we are unable to raise such capital when needed, or on acceptable terms, we may be forced to delay, reduce or eliminate programs, product candidates (including clinical trials), or future commercialization efforts.
•
We are dependent on the success of our product candidates, including CITY-FXI, CITY-RBP4, and CITY-TFR2, and our ongoing and anticipated clinical trials may not be successful.
•
Preclinical and clinical drug development is a lengthy and expensive process, with uncertain timelines and outcomes. If preclinical studies or clinical trials of our product candidates are prolonged or delayed, we may be unable to obtain required regulatory approvals, and therefore be unable to commercialize our product candidates or any of our future product candidates on a timely basis or at all.
•
Regulatory Authorities may not accept data from clinical trials conducted outside the United States, which could require us to conduct additional clinical trials and delay the development or approval of our product candidates.
•
If our clinical trials fail to replicate positive results from earlier preclinical studies or clinical trials conducted by us or third parties, we may be unable to successfully develop, obtain regulatory approval for or commercialize our product candidates.
•
We may encounter substantial delays in the commencement, enrollment or completion of our planned clinical trials or we may fail to demonstrate safety and efficacy to the satisfaction of applicable Regulatory Authorities, which could prevent us from commercializing any product candidates we determine to develop on a timely basis, if at all.
•
If we become involved in patent litigation or other proceedings related to a determination of rights, we could incur substantial costs and expenses, substantial liability for damages or be required to stop our product development and commercialization efforts.
•
Our success is largely based upon our intellectual property and proprietary technologies, and we may be unable to adequately protect and/or enforce our intellectual property.
•
We are dependent on intellectual property that we have in-licensed from third parties, and our failure to comply with our obligations under these licenses could result in loss of rights that are critical to our business.
•
We rely on and expect to continue to rely on third parties to conduct aspects of our research, preclinical studies, clinical protocol development and clinical trials for our programs and product candidates. If these third parties do not perform satisfactorily, comply with regulatory requirements or meet expected deadlines, we may not be able to develop product candidates in a timely or cost-effective manner, or obtain regulatory approval for or commercialize our product candidates and our business could be substantially harmed.
•
We rely on third parties for the supply and manufacture of our product candidates for our research, preclinical, and clinical activities, and may do the same for commercial supplies of our products, if approved. As our pipeline increases and matures, the increased demand for supplies from our manufacturers may increase the risk that we will not have sufficient supply when needed or at an acceptable cost.
•
We depend on limited source suppliers for certain drug substances, drug products, raw materials, samples, components and other materials used in our product candidates. If we are unable to source these supplies on a timely basis, or establish longer-term contracts with our suppliers, we may not be able to complete our clinical trials on time and the development of our product candidates may be delayed.
•
We have in the past entered into, and in the future may enter into, partnership, collaboration and licensing arrangements with third parties to support development of programs and product candidates. If these partnership, collaboration, and licensing arrangements are not successful, our business could be adversely affected.
•
The biopharmaceutical market is intensely competitive. If we or our collaborators are unable to compete effectively with existing drugs, new treatment methods, and new technologies, including with respect to our specific product candidates CITY-FXI, CITY-RBP4, and CITY-TFR2, we may be unable to successfully commercialize any drugs that we develop.
•
There has been no prior public market for our common stock, and an active trading market for our common stock may not develop.
•
The price of our common stock may be volatile and fluctuate substantially, which could result in substantial losses for purchasers of our common stock in this offering.
The summary risk factors described above should be read together with the text of the full risk factors in the section titled “Risk Factors” and the other information set forth in this prospectus, including our audited consolidated financial statements and the related notes, as well as in other documents that we file with the Securities and Exchange Commission (the “SEC”). The risks summarized above or described in full elsewhere in this prospectus are not the only risks that we face. Additional risks and uncertainties not presently known to us, or that we currently deem to be immaterial may also materially adversely affect our business, financial condition, results of operations, and future prospects.
Recent Developments
Certain Preliminary Financial Information (Unaudited)
As of September 30, 2026, we estimate that we had approximately $141.9 million in cash and cash equivalents. This amount is preliminary and unaudited and is subject to completion of our financial closing procedures. This preliminary unaudited financial data has been prepared by, and is the responsibility of, our management. PricewaterhouseCoopers LLP, our independent registered public accounting firm, has not audited, reviewed, examined, compiled, nor applied agreed-upon procedures with respect to the preliminary financial data. Accordingly, PricewaterhouseCoopers LLP does not express an opinion or any other form of assurance with respect thereto. Further, this estimate does not present all information necessary for an understanding of our financial condition and liquidity as of and for the nine months ended September 30, 2026. This estimate should not be viewed as a substitute for financial statements prepared in accordance with accounting principles generally accepted in the United States and is not necessarily indicative of the results to be achieved in any future period. You should not draw any conclusions based on the foregoing estimate and should not place undue reliance on this preliminary estimate. We assume no duty to update this preliminary estimate except as required by law.
Corporate Information
We were incorporated under the laws of the State of Delaware in July 2023. Our principal executive offices are located at 399 Binney Street, Cambridge, Massachusetts 02142, and our telephone number is (857) 357-2024. We have one subsidiary, City Therapeutics Securities Corporation, formed in
August 2024 under the laws of the Commonwealth of Massachusetts. Our website address is www.citytx.com. The information contained in or accessible from our website is not incorporated into this prospectus, and you should not consider it part of this prospectus. We have included our website address in this prospectus solely as an inactive textual reference.
Implications of Being an Emerging Growth Company and a Smaller Reporting Company
We qualify as an “emerging growth company” as defined in the Jumpstart Our Business Startups Act of 2012, as amended (the “JOBS Act”). As an emerging growth company, we may take advantage of specified reduced disclosure and other requirements that are otherwise applicable generally to public companies. These provisions include:
•
being permitted to present only two years of audited financial statements, in addition to any required unaudited interim financial statements, with correspondingly reduced “Management’s Discussion and Analysis of Financial Condition and Results of Operations” disclosure in this prospectus;
•
reduced disclosure about our executive compensation arrangements;
•
not being required to hold advisory votes on executive compensation or to obtain stockholder approval of any golden parachute arrangements not previously approved;
•
an exemption from the auditor attestation requirement in the assessment of our internal control over financial reporting pursuant to Section 404 of the Sarbanes-Oxley Act of 2002 (the “Sarbanes-Oxley Act”); and
•
an exemption from compliance with the requirements of the Public Company Accounting Oversight Board regarding the communication of critical audit matters in the auditor’s report on the financial statements.
We may take advantage of these exemptions for up to five years or such earlier time that we are no longer an emerging growth company. We may choose to take advantage of some but not all of these exemptions. We would cease to be an emerging growth company on the date that is the earliest of (i) the last day of the fiscal year in which we have total annual gross revenues of $1.235 billion or more; (ii) the last day of our fiscal year following the fifth anniversary of the date of the completion of this offering; (iii) the date on which we have issued more than $1.0 billion in non-convertible debt during the previous three years; or (iv) the date on which we are deemed to be a large accelerated filer under the rules of the SEC. We have taken advantage of reduced reporting requirements in this prospectus. Accordingly, the information contained herein may be different from the information you receive from other public companies in which you hold stock. Additionally, the JOBS Act provides that an emerging growth company can take advantage of an extended transition period for complying with new or revised accounting standards. This allows an emerging growth company to delay the adoption of certain accounting standards until those standards would otherwise apply to private companies. We have elected to avail ourselves of this exemption and, therefore, while we are an emerging growth company we will not be subject to new or revised accounting standards at the same time that they become applicable to other public companies that are not emerging growth companies. As a result of this election, our consolidated financial statements may not be comparable to those of other public companies that comply with new or revised accounting pronouncements as of public company effective dates. We may choose to early adopt any new or revised accounting standards whenever such early adoption is permitted for private companies.
We are also a “smaller reporting company,” meaning that the market value of our shares held by non-affiliates plus the proposed aggregate amount of gross proceeds to us as a result of this offering is less than $700 million and our annual revenue was less than $100 million during the most recently completed fiscal year. We may continue to be a smaller reporting company after this offering if either (i) the market value of our shares held by non-affiliates is less than $250 million measured on the last business day of our second fiscal quarter or (ii) our annual revenue was less than $100 million during the most recently completed fiscal year and the market value of our shares held by non-affiliates is less than $700 million measured on the last business day of our second fiscal quarter. If we are a smaller
reporting company at the time we cease to be an emerging growth company, we may continue to rely on exemptions from certain disclosure requirements that are available to smaller reporting companies. Specifically, as a smaller reporting company, we may choose to present only the two most recent fiscal years of audited consolidated financial statements in our Annual Report on Form 10-K and, similar to emerging growth companies, smaller reporting companies have reduced disclosure obligations regarding executive compensation.
THE OFFERING
Common stock offered by us
9,722,222 shares.
Option to purchase additional shares of common stock
We have granted a 30-day option to the underwriters to purchase up to 1,458,333 additional shares of common stock from us at the public offering price, less underwriting discounts and commissions.
Common stock to be outstanding immediately after this offering
50,332,279 shares (or 51,790,612 shares if the underwriters exercise their option to purchase additional shares of common stock in full).
We estimate that the net proceeds from the sale of our common stock in this offering will be approximately $158.3 million (or approximately $182.7 million if the underwriters exercise their option to purchase additional shares of common stock in full), based on the assumed initial public offering price of $18.00 per share, which is the midpoint of the estimated offering price range set forth on the cover page of this prospectus, after deducting underwriting discounts and commissions and estimated offering expenses payable by us.
We currently intend to use the net proceeds from this offering, together with our existing cash and cash equivalents, as follows: approximately $30.0 million to fund the clinical development of CITY-FXI, including our ongoing Phase 1 clinical trial and the initiation of a Phase 2 TKA study; approximately $20.0 million to fund the clinical development of CITY-RBP4, including our Phase 1 clinical trial and preparations for later-stage clinical development for the treatment of Stargardt disease and GA; approximately $40.0 million to fund the clinical development of CITY-TFR2, including an IIT in China and the Phase 1/2 clinical trial, both focused on the treatment of anemia associated with myelofibrosis; approximately $70.0 million to advance our City Platform, including continued investment to advance cityRNA triggers (CITY-O02 and CITY-C02) to the pipeline, as well as additional preclinical product candidates toward IND or IND-equivalent submission and clinical development; and the remainder for working capital and other general corporate purposes. See the section titled “Use of Proceeds” for additional information.
See the section titled “Risk Factors” for a discussion of factors you should carefully consider before deciding whether to invest in our common stock.
Proposed Nasdaq Global Market trading symbol
“CTY”
The number of shares of our common stock that will be outstanding after this offering is based on 40,610,057 shares of our common stock outstanding as of June 30, 2026 (which includes 1,964,421 shares of restricted common stock subject to repurchase or forfeiture), after giving effect to (i) the automatic conversion of all outstanding shares of our convertible preferred stock into an aggregate of 28,848,641 shares of our common stock and (ii) the automatic conversion of the outstanding principal and accrued interest under the convertible promissory note with Biogen (the “Biogen Note”) into an aggregate of 2,133,788 shares of our common stock at a conversion price equal to 85% of the assumed initial public offering price of $18.00 per share, the midpoint of the estimated offering price range set forth on the cover page of this prospectus, in each case immediately prior to the completion of this offering, and excludes:
•
5,404,513 shares of common stock issuable upon exercise of outstanding stock options as of June 30, 2026 under our 2023 Stock Plan, as amended from time to time (the “2023 Plan”), with a weighted-average exercise price of $1.97 per share;
•
988,458 shares of common stock issuable upon exercise of outstanding stock options granted after June 30, 2026 under our 2023 Plan, with a weighted-average exercise price of $8.64 per share;
•
1,482,874 shares of common stock reserved for future issuance as of June 30, 2026 under our 2023 Plan, which will cease to be available for issuance at the time that our 2026 Stock Option and Incentive Plan (the “2026 Plan”) becomes effective;
•
6,516,101 shares of our common stock reserved for future issuance under our 2026 Plan, which will become effective on the date immediately prior to the effectiveness of the registration statement of which this prospectus forms a part, as well as any automatic increases in the number of shares of common stock reserved for future issuance under the 2026 Plan and any shares underlying outstanding stock awards granted under our 2023 Plan that expire or are repurchased, forfeited, cancelled or withheld; and
•
503,322 shares of common stock reserved for future issuance under our 2026 Employee Stock Purchase Plan (the “ESPP”), which will become effective on the date immediately prior to the effectiveness of the registration statement of which this prospectus forms a part, as well as any automatic increases in the number of shares of common stock reserved for future issuance under the ESPP.
Unless otherwise indicated, the information in this prospectus reflects or assumes the following:
•
the automatic conversion of (i) all outstanding shares of our convertible preferred stock into an aggregate of 28,848,641 shares of common stock and (ii) the outstanding principal and accrued interest under the Biogen Note into an aggregate of 2,133,788 shares of our common stock at a conversion price equal to 85% of the assumed initial public offering price of $18.00 per share, the midpoint of the estimated offering price range set forth on the cover page of this prospectus, in each case immediately prior to the completion of this offering;
•
no exercise of the outstanding stock options or restricted common stock described above after June 30, 2026;
•
no exercise of the underwriters’ option to purchase up to an additional 1,458,333 shares of common stock in this offering;
•
a 1.0983-for-one stock split of our common stock, which will become effective prior to the completion of this offering and a corresponding adjustment to the ratio at which our convertible preferred stock will convert into common stock; and
•
the filing and effectiveness of our amended and restated certificate of incorporation (the “amended and restated certificate of incorporation”) immediately prior to the completion of this offering and the effectiveness of our amended and restated bylaws upon the effectiveness of the registration statement of which this prospectus forms a part.
SUMMARY CONSOLIDATED FINANCIAL DATA
The following tables set forth our summary consolidated statements of operations for the years ended December 31, 2024 and 2025 and for the six months ended June 30, 2025 and 2026 and our summary condensed consolidated balance sheet data as of June 30, 2026. The summary consolidated statements of operations data for the years ended December 31, 2024 and 2025 have been derived from our audited consolidated financial statements included elsewhere in this prospectus. The summary condensed consolidated statements of operations data for the six months ended June 30, 2025 and 2026 and summary condensed consolidated balance sheet data as of June 30, 2026 have been derived from our unaudited condensed consolidated financial statements included elsewhere in this prospectus. Our historical results are not necessarily indicative of the results that may be expected for any period in the future, and our interim results are not necessarily indicative of the results that may be expected for the full year or any other period. Our unaudited interim financial statements were prepared on the same basis as our audited financial statements and include, in the opinion of management, all adjustments, consisting of normal recurring adjustments, that are necessary for the fair statement of the financial information set forth in those financial statements. You should read the following summary financial data together with “Management’s Discussion and Analysis of Financial Condition and Results of Operations” and our consolidated financial statements and the related notes included elsewhere in this prospectus. The summary consolidated financial data included in this section are not intended to replace the consolidated financial statements and are qualified in their entirety by our consolidated financial statements and the related notes included elsewhere in this prospectus.
|
(in thousands, except share and per share amounts)
|
|
|
Year ended December 31,
|
|
|
Six Months ended June 30,
|
|
| |
2024
|
|
|
2025
|
|
|
2025
|
|
|
2026
|
|
|
Consolidated Statements of Operations and Comprehensive Loss Data:
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
Collaboration revenue
|
|
|
|
$ |
2,778 |
|
|
|
|
$ |
13,562 |
|
|
|
|
$ |
5,578 |
|
|
|
|
$ |
6,358 |
|
|
|
Operating expenses:
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
Research and development(1)
|
|
|
|
|
25,682 |
|
|
|
|
|
58,420 |
|
|
|
|
|
24,375 |
|
|
|
|
|
46,319 |
|
|
|
General and administrative(2)
|
|
|
|
|
6,186 |
|
|
|
|
|
15,064 |
|
|
|
|
|
7,796 |
|
|
|
|
|
11,170 |
|
|
|
Total operating expenses
|
|
|
|
|
31,868 |
|
|
|
|
|
73,484 |
|
|
|
|
|
32,171 |
|
|
|
|
|
57,489 |
|
|
|
Loss from operations
|
|
|
|
|
(29,090) |
|
|
|
|
|
(59,922) |
|
|
|
|
|
(26,593) |
|
|
|
|
|
(51,131) |
|
|
|
Other income:
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
Interest income
|
|
|
|
|
2,397 |
|
|
|
|
|
4,208 |
|
|
|
|
|
2,064 |
|
|
|
|
|
1,595 |
|
|
|
Interest expense
|
|
|
|
|
(448) |
|
|
|
|
|
(2,257) |
|
|
|
|
|
(646) |
|
|
|
|
|
(1,496) |
|
|
|
Other expense, net
|
|
|
|
|
(695) |
|
|
|
|
|
(534) |
|
|
|
|
|
893 |
|
|
|
|
|
(2,411) |
|
|
|
Total other income, net
|
|
|
|
|
1,254 |
|
|
|
|
|
1,417 |
|
|
|
|
|
2,311 |
|
|
|
|
|
(2,312) |
|
|
|
Net loss and comprehensive loss
|
|
|
|
$ |
(27,836) |
|
|
|
|
$ |
(58,505) |
|
|
|
|
$ |
(24,282) |
|
|
|
|
$ |
(53,443) |
|
|
|
Net loss per share, basic and diluted(3)
|
|
|
|
$ |
(7.45) |
|
|
|
|
$ |
(11.34) |
|
|
|
|
$ |
(5.02) |
|
|
|
|
$ |
(7.53) |
|
|
|
Weighted-average common shares outstanding, basic and diluted(3)
|
|
|
|
|
3,738,335 |
|
|
|
|
|
5,157,781 |
|
|
|
|
|
4,832,880 |
|
|
|
|
|
7,096,572 |
|
|
|
Pro forma net loss per share allocable to common stockholders, basic and diluted (unaudited)(4)
|
|
|
|
|
— |
|
|
|
|
$ |
(1.77) |
|
|
|
|
|
— |
|
|
|
|
$ |
(1.40) |
|
|
|
Pro forma weighted average common shares outstanding, basic and diluted (unaudited)(4)
|
|
|
|
|
— |
|
|
|
|
|
36,121,584 |
|
|
|
|
|
— |
|
|
|
|
|
38,079,000 |
|
|
(1)
Includes related party amounts of $3.6 million and $5.3 million for research and development expenses for the years ended December 31, 2024 and 2025, respectively, and $1.7 million and $3.8 million for the six months ended June 30, 2025 and 2026, respectively. See Note 17 — “Related Parties” to our audited consolidated financial statements and Note 17 — “Related Parties” to our unaudited condensed consolidated financial statements included elsewhere in this prospectus.
(2)
Includes related party amounts of $0.7 million and $1.1 million for general and administrative expenses for the years ended December 31, 2024 and 2025, respectively, and $0.6 million and $0.7 million for the six months ended June 30, 2025 and 2026, respectively. See Note 17 — “Related Parties” to our audited consolidated financial statements and Note 17 — “Related Parties” to our unaudited condensed consolidated financial statements included elsewhere in this prospectus.
(3)
See Notes 2 and 16 to our consolidated financial statements and Note 16 to our unaudited condensed consolidated financial statements included elsewhere in this prospectus for a description of the method used to calculate net loss per share, basic and diluted.
(4)
Pro forma net loss and comprehensive loss for the year ended December 31, 2025 has been adjusted to reflect a $5.3 million loss from the remeasurement of the Biogen Note to fair value immediately prior to its assumed conversion as of January 1, 2025. There was no adjustment to the pro forma net loss and comprehensive loss for the six months ended June 30, 2026. Pro forma net loss per share allocable to common stockholders, basic and diluted for the year ended December 31, 2025 and the six months ended June 30, 2026 was calculated using the weighted average number of shares of common stock outstanding, including the pro forma effect of (i) the automatic conversion of all outstanding shares of our convertible preferred stock into an aggregate of 28,848,641 shares of our common stock, and (ii) the automatic conversion of the outstanding principal and accrued interest under the Biogen Note into 2,133,788 shares of our common stock, at a conversion price equal to 85% of the assumed initial public offering price of $18.00 per share, the midpoint of the estimated offering price range set forth on the cover page of this prospectus, in each case as if such conversion had occurred as of January 1, 2025. Pro forma net loss per share allocable to common stockholders, basic and diluted does not include the effect of the shares expected to be sold in this offering. Upon the completion of this offering, if the actual initial public offering price differs from the assumed initial public offering price of $18.00 per share, the midpoint of the estimated offering price range set forth on the cover page of this prospectus, the number of shares of common stock issuable upon conversion of the Biogen Note will differ from the amounts reflected above. A $1.00 decrease in the assumed initial public offering price of $18.00 per share would increase the number of shares of common stock issuable upon conversion of the outstanding principal and accrued interest under the Biogen Note by an aggregate of 125,517 shares. A $1.00 increase in the assumed initial public offering price of $18.00 per share would decrease the number of shares of common stock issuable upon such conversion by an aggregate of 112,304 shares. The actual number of shares of common stock issuable upon conversion will be based on the outstanding principal and accrued interest under the Biogen Note and the actual initial public offering price.
| |
|
|
As of June 30, 2026
|
|
|
(in thousands)
|
|
|
Actual
|
|
|
Pro Forma(1)
|
|
|
Pro Forma As
Adjusted(2)
|
|
|
Consolidated Balance Sheet Data:
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
Cash, cash equivalents, and restricted cash
|
|
|
|
$ |
172,109 |
|
|
|
|
$ |
172,109 |
|
|
|
|
$ |
330,583 |
|
|
|
Working capital(3)
|
|
|
|
|
110,931 |
|
|
|
|
|
144,020 |
|
|
|
|
|
303,165 |
|
|
|
Total assets
|
|
|
|
|
208,611 |
|
|
|
|
|
208,611 |
|
|
|
|
|
366,190 |
|
|
|
Convertible notes
|
|
|
|
|
33,089 |
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
Total liabilities
|
|
|
|
|
84,272 |
|
|
|
|
|
51,183 |
|
|
|
|
|
50,512 |
|
|
|
Convertible preferred stock
|
|
|
|
|
256,056 |
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
Total stockholders’ (deficit) equity
|
|
|
|
|
(131,717) |
|
|
|
|
|
157,428 |
|
|
|
|
|
315,678 |
|
|
(1)
Pro forma amounts give effect to (i) the automatic conversion of all outstanding shares of our convertible preferred stock into an aggregate of 28,848,641 shares of our common stock immediately prior to the completion of this offering, (ii) the automatic conversion of the outstanding principal and accrued interest under the Biogen Note into an aggregate of 2,133,788 shares of our common stock immediately prior to the completion of this offering, at a conversion price equal to 85% of the assumed initial public offering price of $18.00 per share, the midpoint of the estimated offering price range listed on the cover page of this prospectus, including a change in the fair value of the Biogen Note liability of $5.3 million based on the assumed initial public offering price of $18.00 per share, which is the midpoint of the estimated offering price range set forth on the cover page of this prospectus, and (iii) the filing and effectiveness of our amended and restated certificate of incorporation, which will occur immediately prior to the completion of this offering.
(2)
Pro forma as adjusted amounts give effect to (i) the pro forma adjustments set forth in footnote (1) above and (ii) the issuance and sale of 9,722,222 shares of our common stock in this offering at the initial public offering price of $18.00 per share, the midpoint of the price range set forth on the cover page of this prospectus, after deducting the underwriting discounts and commissions and estimated offering expenses payable by us. This pro forma as adjusted information is illustrative only and will depend on the actual initial public offering price and other terms of this offering determined at pricing. Each $1.00 increase or decrease in the assumed initial public offering price of $18.00 per share would increase or decrease, as applicable, the pro forma as adjusted amount of each of our cash, cash equivalents, and restricted cash, working capital, total assets and total stockholders’ (deficit) equity by approximately $9.0 million, assuming that the number of shares offered by us, as set forth on the cover page of this prospectus, remains the same and after deducting the underwriting discounts and commissions and estimated offering expenses payable by us. Each increase or decrease of 1.0 million shares in the number of shares offered by us at the assumed initial public offering price of $18.00 per share would increase or decrease, as applicable, the pro forma as adjusted amounts of each of our cash, cash equivalents, and restricted cash, working capital, total assets and total stockholders’ (deficit) equity by approximately $16.7 million, assuming no change in the assumed initial public offering price per share, and after deducting the underwriting discounts and commissions and estimated offering expenses payable by us.
(3)
We define working capital as current assets less current liabilities. See our unaudited condensed consolidated financial statements and the related notes included elsewhere in this prospectus for further details regarding our current assets and current liabilities.
RISK FACTORS
Investing in our common stock involves a high degree of risk. You should carefully consider the following risks and uncertainties, together with all other information in this prospectus, including our consolidated financial statements and related notes and “Management’s Discussion and Analysis of Financial Condition and Results of Operations,” before investing in our common stock. Any of the risk factors we describe below could adversely affect our business, financial condition or results of operations. The market price of our common stock could decline if one or more of these risks or uncertainties actually occur, causing you to lose all or part of the money you paid to buy our common stock. Additional risks that we currently do not know about or that we currently believe to be immaterial may also impair our business. Certain statements below are forward-looking statements. See the section titled “Cautionary Note Regarding Forward-Looking Statements” appearing elsewhere in this prospectus.
Risks Related to Our Limited Operating History, Financial Position and Need for Additional Capital
We are a clinical-stage biotechnology company with a limited operating history and no products approved by regulators for commercial sale, which may make it difficult to evaluate our current and future business prospects.
We are a clinical-stage biotechnology company with a limited operating history, which may make it difficult to evaluate the success of our business to date and assess our future viability. Since our inception in July 2023, we have devoted substantially all of our efforts and financial resources to organizing and staffing our company, business planning, establishing our intellectual property portfolio, raising capital, advancing development of our portfolio of programs and platform, including identifying and developing our product candidates, conducting research and preclinical studies and IND-enabling studies, and initiating and conducting clinical trials, and providing general and administrative support to our operations. All of our programs and product candidates are still in the research, preclinical development or clinical development stages. We have not yet demonstrated our ability to successfully complete any clinical trials, obtain regulatory approvals, manufacture a commercial-scale product or arrange for a third party to do so on our behalf, or conduct sales, marketing and distribution activities necessary for successful product commercialization. Additionally, we expect our financial condition and operating results to continue to fluctuate significantly from period to period due to a variety of factors, many of which are beyond our control. Consequently, any predictions made about our future success or viability may not be as accurate as they could be if we had a longer operating history.
We have no products approved for commercial sale and we can provide no assurance that we will obtain regulatory approvals to market and sell any products in the future. We therefore have never generated any revenue from product sales, and we do not expect to generate any revenue from product sales in the foreseeable future. Biopharmaceutical product development is a highly speculative undertaking and involves a substantial degree of risk. If we do not address these risks and difficulties successfully, our business will suffer.
We have incurred significant losses since our inception, including net losses of $27.8 million and $58.5 million for the years ended December 31, 2024 and 2025, respectively, and $24.3 million and $53.4 million for the six months ended June 30, 2025 and 2026, respectively. As of June 30, 2026 we had an accumulated deficit of $142.5 million. We anticipate that we will continue to incur significant losses for the foreseeable future.
We have no products approved for commercial sale and have not generated any revenue from product sales to date. We will continue to incur significant research and development and other expenses related to our programs, product candidates and ongoing operations. As a result, we are not profitable and have incurred losses in each period since our inception. Net losses and negative cash flows have had, and will continue to have, an adverse effect on our stockholders’ equity (deficit) and working capital. We have incurred net losses in each year since our inception in 2023, including net losses of $27.8 million and $58.5 million during the years ended December 31, 2024 and 2025, respectively, and
$24.3 million and $53.4 million for the six months ended June 30, 2025 and 2026, respectively. As of June 30, 2026, we had an accumulated deficit of $142.5 million.
To date, we have financed our operations primarily through the sale and issuance of our convertible preferred stock, proceeds from issuances of convertible notes to our collaboration partners, and upfront and milestone payments under our collaboration and license arrangements. The amount of our future net losses will depend, in part, on the rate of our future expenditures and our ability to obtain funding through public or private equity or debt financings, government or other third-party grants, collaboration and licensing arrangements, asset sales, royalty financings, or a combination of these approaches. We have not completed clinical trials for any of our product candidates, and it will be several years, if ever, before we or our collaboration partners have a product candidate ready for commercialization. Even if we or our collaboration partners obtain regulatory approval to market a product, our future revenues will depend upon the size of any markets in which such product has received approval, and our ability to achieve sufficient market acceptance, reimbursement from third-party payors, and adequate market share in those markets. We may never achieve profitability.
We expect to continue to incur significant expenses and increasing operating losses for the foreseeable future. We anticipate that our expenses will increase substantially if and as we:
•
advance our product candidates through preclinical and clinical development, including continuing the Phase 1 clinical trial for CITY-FXI and advancing other programs through preclinical and clinical development;
•
seek regulatory approvals from the FDA, European Medicines Agency (“EMA”), and/or other foreign comparable regulatory authorities (collectively, the “Regulatory Authorities”) for our product candidates that successfully complete clinical trials;
•
hire additional clinical, quality control, medical, scientific, and other technical personnel to support the clinical development of our product candidates;
•
increase our headcount as we expand our research and development organization, market development and pre-commercial planning activities;
•
undertake activities to establish sales, marketing, and distribution capabilities in preparation for commercialization for our product candidates, as applicable, including hiring additional personnel to support such operations;
•
advance our preclinical-stage product candidates into clinical development;
•
seek to identify, acquire, and further develop additional product candidates where we can leverage our City Platform, our proprietary, integrated platform, for the development of next-generation RNAi therapeutics, including through business development efforts to invest in or in-license other technologies or product candidates;
•
maintain, expand, and protect our intellectual property portfolio;
•
make milestone, royalty, or other payments due under our collaboration and license agreements, and any future in-license or collaboration agreements with third parties; and incur additional legal, audit, regulatory, tax and other expenses associated with operating as a public company.
Biopharmaceutical product development entails substantial upfront capital expenditures and significant risk that a product candidate will fail to demonstrate adequate efficacy or an acceptable safety profile, gain regulatory approval, secure market access and reimbursement and become commercially viable, and therefore any investment in us is highly speculative. Accordingly, before making an investment in us, you should consider our prospects, factoring in the costs, uncertainties, delays and difficulties frequently encountered by companies in clinical development, especially clinical-stage biopharmaceutical companies such as ours. Any predictions you make about our future success or viability may not be as accurate as they would otherwise be if we had a longer operating history or a history of successfully developing and commercializing biopharmaceutical products. We may encounter unforeseen expenses, difficulties, complications, delays and other known or unknown factors in achieving our business objectives.
The development of our City Platform and the development and potential commercialization of our product candidates will require substantial additional capital to fund expenses. We have entered into collaboration agreements for certain programs and product candidates and may decide to collaborate for the future development and potential commercialization of other product candidates. For example, we are party to collaboration agreements with Bausch + Lomb and Biogen pursuant to which we conduct research activities to discover RNAi molecules and related product candidates, while our collaboration partners are primarily responsible for development activities. These agreements contemplate ongoing funding commitments by the parties, including our funding of certain research activities and, in the case of our collaboration with Bausch + Lomb, reimbursement by Bausch + Lomb of specified research costs. However, we cannot guarantee that either we or our collaboration partners will have sufficient funds to support the research and development activities contemplated by these agreements. If we elect to increase our expenditures to fund development or commercialization activities on our own, we may need to obtain additional capital, which may not be available to us on acceptable terms or at all. If we do not have sufficient funds, we may not be able to further develop our programs and product candidates or bring them to market and generate product revenue.
Additionally, our expenses could increase beyond our expectations if we are required by Regulatory Authorities to perform clinical trials in addition to those that we currently have planned for, or if there are any delays in establishing appropriate manufacturing arrangements for our product candidates or in completing clinical trials for our product candidates.
We have never generated any revenue from sales of drug products, and our ability to generate revenue from sales of drug products and become profitable depends significantly on our success in a number of factors.
We have no drug products approved for commercial sale, have not generated any revenue from drug product sales, and do not anticipate generating any revenue from drug product sales until sometime after we have received regulatory approval, if obtained at all, for the commercial sale of our product candidates. Our ability to generate revenue and achieve profitability depends significantly on our success in many factors, including but not limited to: completing research regarding, and preclinical and clinical development of, our product candidates; obtaining regulatory approvals and marketing authorizations for product candidates for which we complete clinical studies; developing a sustainable and scalable manufacturing process for our product candidates, including establishing and maintaining commercially viable supply relationships with third parties and establishing our own manufacturing capabilities and infrastructure; launching and commercializing product candidates for which we obtain regulatory approvals and marketing authorizations; addressing any competing technological and market developments; identifying, assessing, acquiring and/or developing new product candidates, intellectual property and technologies; negotiating favorable terms in any collaboration, licensing, or other arrangements into which we may enter; maintaining, protecting, expanding and enforcing our portfolio of intellectual property rights, including patents, trademarks, trade secrets, and know-how; or attracting, hiring, and retaining qualified personnel.
Even if one or more of the product candidates that we develop is approved for commercial sale, we anticipate incurring significant costs associated with commercializing any approved product candidate. Our expenses could increase beyond expectations if we are required by Regulatory Authorities to change our manufacturing processes or assays, or to perform clinical, preclinical, or other types of studies in addition to those we currently anticipate. If we are successful in obtaining regulatory approvals to market one or more of our product candidates, our revenue will be dependent, in part, upon the size of the market for the relevant product, the accepted price for the product to be paid with out-of-pocket expenses and the ability to get reimbursement for any amount. If the number of patients with our addressable disease is not as significant as we estimate, the indication approved by Regulatory Authorities is narrower than we expect, or the reasonably accepted population for treatment is narrowed by competition, physician choice or treatment guidelines, we may not generate significant revenue from sales of such products, even if approved. If we are not able to generate revenue from the sale of any approved products, we may never become profitable.
Even if this offering is successful, we will require substantial additional capital to finance our operations in the future. If we are unable to raise such capital when needed, or on acceptable terms, we may be forced to delay, reduce or eliminate programs, product candidates (including clinical trials), or future commercialization efforts.
Developing biopharmaceutical products, including conducting preclinical studies and clinical trials, is a time-consuming, expensive and uncertain process that takes years to complete. We expect our expenses to continue to increase in connection with our ongoing activities, particularly as we conduct clinical trials of, and seek regulatory and marketing approval for, our product candidates. Even if our current or future product candidates are approved for commercial sale, we anticipate incurring significant costs associated with commercializing any approved product candidate. To date, we have funded our operations principally through the sale and issuance of our convertible preferred stock and convertible notes, as well as capital received from our collaboration and license arrangements. We expect our expenses to increase in connection with our ongoing activities, particularly as we continue the clinical and preclinical development of our product candidates, continue to identify additional targets using our City Platform, commence additional preclinical studies and clinical trials, and continue to identify and develop additional product candidates either through internal development or through acquisitions or in-licensing product candidates.
As of June 30, 2026, we had cash, cash equivalents, and restricted cash of $172.1 million. Based on our current operating plan, we believe that our existing cash and cash equivalents, together with the net proceeds from this offering, will be sufficient to meet our working capital and capital expenditure needs into late 2028. We have based this estimate on assumptions that may prove to be wrong, and we could utilize our available capital resources sooner than we expect. We may also raise additional financing on an opportunistic basis in the future. For example, we may seek additional capital due to favorable market conditions or strategic considerations even if we believe we have sufficient funds for our current or future operating plans. Attempting to secure additional financing may divert our management from our day-to-day activities, which may adversely affect our ability to develop our product candidates. Our future capital requirements will depend on many factors, including but not limited to:
•
the scope, timing, progress, costs, and results of discovery, preclinical development and clinical trials for our current or future product candidates and the effectiveness of our City Platform;
•
the number of clinical trials required for regulatory approval of our current or future product candidates;
•
the costs, timing, and outcome of regulatory review of any of our current or future product candidates;
•
the costs associated with acquiring or in-licensing additional product candidates, technologies or assets, including the timing and amount of any milestones, royalties, or other payments due in connection with our acquisitions and in-licenses, as applicable;
•
the cost of manufacturing clinical and commercial supplies of our current or future product candidates;
•
the costs and timing of preparing, filing, and prosecuting patent applications, maintaining and enforcing our intellectual property rights and defending any intellectual property-related claims, including any claims by third parties that we are infringing upon their intellectual property rights;
•
our ability to maintain existing, and establish new, strategic collaborations, licensing or other arrangements on favorable terms, if at all, and the financial terms of any such agreements, including the timing and amount of any future milestone, royalty or other payments due to or payable by us thereunder and the extent to which we are obligated to reimburse, or entitled to reimbursement of, clinical trial costs under any such agreements, if any;
•
the costs and timing of future commercialization activities, including manufacturing, marketing, sales, and distribution, for any of our product candidates for which we receive marketing approval;
•
the revenue, if any, received from commercial sales of our product candidates for which we receive marketing approval;
•
expenses to attract, hire, and retain skilled personnel;
•
the costs of operating as a public company;
•
our ability to establish a commercially viable pricing structure and obtain approval for coverage and adequate reimbursement from third-party and government payors;
•
the effect of macroeconomic trends including inflationary pressures, tariffs, and fluctuating interest rates;
•
addressing any potential supply chain interruptions or delays;
•
the effect of competing technological and market developments; and
•
the extent to which we acquire or invest in businesses, products, and technologies.
We expect our expenses to continue to increase in connection with our ongoing activities, particularly as we identify, continue the research and development of, initiate preclinical studies and clinical trials of, and seek marketing approval for, our current or future product candidates, and we may never generate the necessary data or results required to obtain regulatory approval and achieve product sales. In addition, our current or future product candidates, if approved, may not achieve commercial success. Our commercial revenues, if any, will be derived from sales of products that we do not expect to be commercially available for many years, if at all. Accordingly, we will need to continue to rely on additional financing to achieve our business objectives. Adequate additional financing may not be available to us on acceptable terms, or at all. In addition, we may seek additional capital due to favorable market conditions or strategic considerations even if we believe we have sufficient funds for our current or future operating plans. To the extent that we raise additional capital through the sale of equity or convertible debt securities, your ownership interest will be diluted, and the terms of these securities may include liquidation or other preferences that adversely affect your rights as a common stockholder. Debt financing, if available, may involve agreements that include covenants limiting or restricting our ability to take specific actions, such as incurring additional debt, making capital expenditures, declaring dividends, and possibly other restrictions.
Any additional fundraising efforts may divert our management from their day-to-day activities, which may adversely affect our ability to develop and commercialize our product candidates. We have no committed sources of additional capital and, if we are unable to raise additional capital in sufficient amounts or on terms acceptable to us, we may have to significantly delay, scale back or discontinue the development or commercialization of our current or future product candidates or other research and development initiatives. Without sufficient funding, our license agreements and any future collaboration agreements may also be terminated if we are unable to meet the payment or other obligations under such agreements.
If we are unable to raise additional funds through equity or debt financings when needed, we may be required to delay, limit, reduce, or terminate our product development or future commercialization efforts or grant rights to develop and market product candidates that we would otherwise prefer to develop and market ourselves. Additionally, if we raise funds through additional collaborations, strategic alliances, or licensing arrangements with third parties, we may have to relinquish valuable rights to our technologies, future revenue streams, research programs, product candidates we develop, or we may have to grant licenses on terms that may not be favorable to us and/or that may reduce the value of our common stock.
Risks Related to Discovery, Development, Regulatory Approval and Potential Commercialization of Our Current or Future Product Candidates
The approach we are taking to discover and develop certain of our product candidates is novel and may never lead to marketable products.
We have concentrated our research and development efforts on RNAi technology, and our future success depends on the successful development of this technology and products based on our City Platform.
The scientific discoveries that form the basis for our efforts to develop product candidates based on our City Platform are relatively new. The scientific evidence to support the feasibility of developing product candidates based on these discoveries is both preliminary and limited. If we do not successfully develop and commercialize product candidates based upon our technological approach, we may not become profitable, and the value of our common stock may decline.
Further, our focus solely on RNAi technology for developing drugs as opposed to multiple, more proven technologies for drug development increases the risks associated with the ownership of our securities. If we are not successful in developing any product candidates using RNAi technology, we may be required to change the scope and direction of our product development activities. In that case, we may not be able to identify and successfully implement an alternative product development strategy.
We are early in our development efforts. We have only recently initiated early clinical studies, and as a result it will be years before we commercialize a product candidate, if ever. If we are unable to identify and advance product candidates through preclinical studies and clinical trials, obtain marketing approval and ultimately commercialize them, or experience significant delays in doing so, our business will be materially harmed.
The success of our business depends primarily upon our ability to identify, develop and commercialize product candidates. We are early in our development efforts and our lead product candidate, CITY-FXI, is in early clinical development and our other product candidates and development programs are in preclinical development or drug discovery stages. We have invested substantially all of our research efforts to date in developing our City Platform, identifying potential product candidates and conducting preclinical and early clinical studies. As an organization, we have limited experience in conducting and managing clinical trials necessary to obtain regulatory approvals, and we may be unable to do so for our current or future product candidates. While we have successfully completed IND- or IND‑equivalent studies for CITY-FXI and have initiated a Phase 1 clinical trial to assess preliminary safety, tolerability, pharmacokinetic, and pharmacodynamic properties of CITY-FXI, we have not yet completed any clinical trials for CITY-FXI or any of our other product candidates to date. Additionally, we have a portfolio of current product candidates that are in earlier stages of discovery or preclinical development and may never advance to clinical-stage development. If we are able to advance these other product candidates into clinical development, we do not have experience managing multiple clinical trials simultaneously, working with global clinical trials, or working in multiple different disease indications. Our ability to achieve and sustain profitability depends on obtaining regulatory approvals for, and successfully commercializing our product candidates, either alone or with third parties, and we cannot guarantee you that we will ever obtain regulatory approval for any of our product candidates. Before obtaining regulatory approval for the commercial distribution of our product candidates, we must conduct extensive preclinical tests and clinical trials to demonstrate the safety and efficacy in humans of our product candidates.
We may not have the financial resources to continue development of, or the ability to enter into new collaborations for, a product candidate if we experience any issues that delay or prevent regulatory approval of, or our ability to commercialize, product candidates, including:
•
preclinical study results may show the product candidate to be less effective than desired or to have harmful or problematic side effects;
•
negative or inconclusive results from our clinical trials or the clinical trials of others for product candidates similar to ours, leading to a decision or requirement to conduct additional preclinical testing or clinical trials or abandon a program candidate;
•
product-related side effects experienced by patients in our clinical trials or by individuals using products similar to our product candidates;
•
our third-party manufacturers’ inability to successfully manufacture our product candidates;
•
inability of any third-party contract manufacturer to scale up manufacturing of our product candidates and those of our collaborators to supply the needs of clinical trials or commercial sales;
•
delays in submitting INDs, clinical trial authorization applications or other comparable foreign applications or delays or failures in obtaining the necessary approvals from regulators to commence a clinical trial, or a suspension or termination of a clinical trial once commenced;
•
preclinical studies conducted outside of the United States may be affected by tariffs or import/export restrictions imposed by the United States or other foreign governments;
•
conditions imposed by Regulatory Authorities regarding the scope or design of our clinical trials;
•
delays in enrolling patients in our clinical trials;
•
high drop-out rates of our clinical trial patients;
•
inadequate supply or quality of product candidate components or materials or other supplies necessary for the conduct of our clinical trials;
•
inability to obtain alternative sources of supply for which we have a single source for product candidate components or materials;
•
greater than anticipated costs of our clinical trials;
•
manufacturing costs, formulation issues, pricing or reimbursement issues, or other factors that no longer make a product candidate economically feasible;
•
harmful or undesirable side effects or inability of our product candidates to meet efficacy endpoints during clinical trials;
•
failure to demonstrate a benefit-risk profile acceptable to Regulatory Authorities;
•
unfavorable Regulatory Authority inspection and review of any of the clinical trial sites or manufacturing facilities used in the testing and manufacture of any of our product candidates;
•
failure of our third-party contractors or investigators to comply with regulatory requirements or otherwise meet their contractual obligations in a timely manner, or at all;
•
delays and changes in regulatory requirements, policy and guidelines, including the imposition of additional regulatory oversight around clinical testing generally or with respect to our technology in particular; or
•
varying interpretations of our data by Regulatory Authorities.
Our inability to complete development of, or commercialize our product candidates, or significant delays in doing so due to one or more of these factors, could have a material and adverse effect on our business, financial condition, results of operations and prospects.
Our approach to the engineering and development of our programs is unproven, and we may not be successful in our efforts to identify and develop any programs and product candidates of commercial value by leveraging our City Platform.
Our City Platform and approach to RNAi trigger engineering and product candidate development utilize, among other things, our proprietary cityRNA trigger engineering capabilities and proprietary AI/ML molecular-design capabilities to discover, design and engineer RNAi trigger molecules and product candidates. Because our approach is proprietary and incorporates novel technologies, including our cityRNA trigger molecules, the cost and time required to discover and develop our product candidates can be difficult to predict, and our efforts may not result in the discovery and development of commercially viable product candidates.
Any product candidate discovery and development efforts conducted using our City Platform may not be successful in identifying product candidates that have commercial value or therapeutic utility. Our City Platform may initially show promise in identifying, designing or engineering RNAi trigger molecules and product candidates, yet fail to yield viable product candidates for preclinical development, clinical development or potential commercialization for a number of reasons, including:
•
research programs to identify and optimize new RNAi trigger molecules and product candidates will require substantial technical, financial and human resources, and we may be unsuccessful in our efforts to identify or advance suitable product candidates. If we are unable to identify suitable product candidates for preclinical and clinical development, our ability to expand our pipeline and generate future revenues could be adversely affected;
•
City-engineered product candidates discovered using our City Platform may not demonstrate the expected potency, efficacy, safety, tolerability, pharmacokinetic or pharmacodynamic properties in preclinical studies or clinical trials, including because they may exhibit different biological, chemical or pharmacological properties in humans than they do in laboratory studies or otherwise interact with human biological systems in unforeseen, ineffective or potentially harmful ways;
•
our proprietary AI/ML molecular-design capabilities and the datasets on which they are based may not accurately predict the activity, potency or other characteristics of RNAi trigger molecules or otherwise improve our ability to identify or optimize product candidates, and any advantages demonstrated in laboratory testing may not translate into successful product candidate development;
•
product candidates identified through our City Platform may, on further study, be shown to have harmful side effects, off-target activity or other characteristics that indicate they are unlikely to receive regulatory approval or achieve market acceptance;
•
competitors may develop alternative RNAi technologies, delivery approaches or other therapies that render our product candidates noncompetitive or less attractive; or
•
a potential product candidate may not be capable of being manufactured reliably, at commercial scale or at an acceptable cost.
In addition, we may seek to identify and develop product candidates based on novel RNAi trigger molecules, delivery technologies, targets and other approaches that are unproven. In particular, our cityRNA trigger molecules represent a novel class of RNAi trigger molecules, and there is no clinical experience with product candidates incorporating these trigger molecules. If our activities fail to identify suitable RNAi trigger molecules, targets or other technologies for product candidate development, or if such technologies prove to be unsuitable for treating human disease, we may not be able to develop viable additional product candidates. We and our existing or future collaborators may never receive regulatory approval to market and commercialize any product candidate. Even if we or an existing or future collaborator obtains regulatory approval, the approval may be for indications or patient populations that are not as broad as we intended or desired or may require labeling that includes significant use or distribution restrictions or safety warnings. If the product candidates resulting from our City Platform prove to be ineffective, unsafe or commercially unviable, our City Platform and product candidates would have little, if any, value, which could have a material adverse effect on our business, financial condition, results of operations and prospects.
Preclinical and clinical drug development is a lengthy and expensive process, with uncertain timelines and outcomes. If preclinical studies or clinical trials of our product candidates are prolonged or delayed, we may be unable to obtain required regulatory approvals, and therefore be unable to commercialize our product candidates or any of our future product candidates on a timely basis or at all.
Successful development of pharmaceutical products involves a lengthy and expensive process, is highly uncertain, and is dependent on numerous factors, many of which are beyond our control. Product candidates that appear promising in the early phases of development may fail to reach the market for several reasons, including:
•
clinical trial results may show the product candidates to be less effective than expected (for example, a clinical trial could fail to meet its primary or key secondary endpoint(s)) or have an unacceptable safety or tolerability profile;
•
failure to receive the necessary regulatory approvals or a delay in receiving such approvals, which, among other things, may be caused by patients who fail the trial screening process, slow enrollment in clinical trials, patients dropping out of trials, patients lost to follow-up, length of time to achieve trial endpoints, additional time requirements for data analysis or New Drug Application (“NDA”) or similar foreign application preparation, discussions with Regulatory Authorities, including Regulatory Authorities requesting additional preclinical or clinical data (such as long-term toxicology studies), or encountering unexpected safety or manufacturing issues;
•
preclinical study results may show the product candidate to be less effective than desired or to have harmful on-target or off-target side effects;
•
imposition of extensive post-marketing approval requirements; or
•
the proprietary rights of others and their competing products and technologies that may prevent our product candidates from being commercialized.
Furthermore, the length of time necessary to complete clinical trials and submit an application for regulatory approval for a final decision by a regulatory authority varies significantly from one product candidate to the next and from one country or jurisdiction to the next and may be difficult to predict. Even if we are successful in obtaining regulatory approval, commercial success of any approved products will also depend in large part on the availability of coverage and adequate reimbursement from third-party payors, including government payors such as the Medicare and Medicaid programs and managed care organizations in the United States or country-specific governmental organizations in foreign countries, which may be affected by existing and future healthcare reform measures designed to reduce the cost of healthcare. Third-party payors could require us to conduct additional studies, including post-marketing studies related to the cost effectiveness of a product, to qualify for reimbursement, which could be costly and divert our resources. If government and other healthcare payors were not to provide coverage and adequate reimbursement for our products once approved, market acceptance and commercial success would be reduced. Even if we are able to obtain coverage and adequate reimbursement for our products once approved, there may be features or characteristics of our products, such as dose preparation requirements, that prevent our products from achieving market acceptance by the healthcare or patient communities.
In addition, if any of our product candidates receive marketing approval, we will be subject to significant regulatory obligations regarding the submission of safety and other post-marketing information and reports and registration, and will need to continue to comply (or ensure that our third-party providers comply) with current Good Manufacturing Practice (“cGMPs”) and Good Clinical Practice (“GCPs”) for any clinical trials that we conduct post-approval. In addition, there is always the risk that we, a Regulatory Authority or a third party might identify previously unknown problems with a product post-approval, such as adverse events (“AEs”) of unanticipated severity or frequency. Compliance with these requirements is costly, and any failure to comply or other issues with our product candidates post-approval could adversely affect our business, financial condition and results of operations.
We may encounter substantial delays in the commencement, enrollment or completion of our planned clinical trials or we may fail to demonstrate safety and efficacy to the satisfaction of applicable Regulatory Authorities, which could prevent us from commercializing any product candidates we determine to develop on a timely basis, if at all.
The risk of failure in developing product candidates is high. It is impossible to predict when or if any product candidate would prove effective or safe in humans or will receive regulatory approval. Before obtaining marketing approval from applicable Regulatory Authorities for the sale of any product candidate, we must complete preclinical development, submit an IND or comparable foreign application to permit initiation of clinical studies, and then conduct extensive clinical trials to demonstrate the safety and efficacy of product candidates in humans. We have not yet completed a clinical trial of any product candidate.
Before we can commence clinical trials for a product candidate, we must complete extensive preclinical testing and studies that support our INDs and other regulatory filings. We cannot be certain
of the timely identification of a product candidate or the successful completion or outcome of our preclinical testing and studies and cannot predict whether Regulatory Authorities will accept our proposed clinical programs or whether the outcome of our preclinical testing and studies will ultimately support the further development of any product candidates. Conducting preclinical testing is a lengthy, time-consuming and expensive process. The length of time may vary substantially according to the type, complexity and novelty of the program, and often can be several years or more per program. As a result, we cannot be sure that we will be able to submit INDs or other comparable foreign regulatory submissions for our preclinical programs on the timelines we expect, if at all, and we cannot be sure that submission of INDs will result in the FDA, competent authorities of Member States of the European Union (the “EU Member States”), or other comparable foreign regulatory filings will result in Regulatory Authorities allowing clinical trials to begin.
Furthermore, product candidates are subject to continued preclinical safety studies, which may be conducted concurrently with our clinical testing. The outcomes of these safety studies may delay the launch of or enrollment in future clinical trials and could impact our ability to continue to conduct our clinical trials.
Clinical testing is expensive, is difficult to design and implement, can take many years to complete and is uncertain as to outcome. We cannot guarantee that any clinical trials will be conducted as planned or completed on schedule, or at all. A failure of one or more clinical trials can occur at any stage of testing, which may result from a multitude of factors, including, but not limited to, flaws in trial design, dose selection issues, patient enrollment criteria and failure to demonstrate favorable safety or efficacy traits.
Other events that may prevent successful enrollment, initiation or timely completion of clinical development include:
•
we may be unable to generate sufficient preclinical, toxicology or other in vivo or in vitro data to support the initiation of clinical trials;
•
delays in reaching a consensus with applicable Regulatory Authorities on trial design or implementation;
•
delays in obtaining regulatory authorization to commence a clinical trial;
•
delays in reaching agreement on acceptable terms with prospective Contract Research Organizations (“CROs”), other vendors, or clinical trial sites, the terms of which can be subject to extensive negotiation and may vary significantly among different vendors and trial sites;
•
delays in obtaining approval from one or more institutional review boards (“IRB”) or ethics committees refusing to approve, suspending or terminating the trial at an investigational site, precluding enrollment of additional participants, or withdrawing their approval of the trial;
•
delays in recruiting suitable patients to participate in our ongoing and planned clinical trials;
•
changes to the clinical trial protocol;
•
clinical sites deviating from trial protocol or dropping out of a trial;
•
delays in manufacturing sufficient quantities of our product candidates for use in clinical trials, or delays in sufficiently developing, characterizing or controlling a manufacturing process suitable for clinical trials;
•
delays in having our product candidates being shipped on time, clearing customs and arriving at clinical trial sites intact;
•
delays in having patients complete participation in a trial or return for post-treatment follow-up;
•
participants choosing an alternative treatment for the indication for which we are developing our product candidates, or participating in competing clinical trials;
•
lack of adequate funding to continue a clinical trial;
•
occurrence of AEs or serious adverse events (“SAEs”) associated with the product candidate that are viewed to outweigh its potential benefits;
•
occurrence of SAEs in clinical trials of the same class of agents conducted by other companies;
•
imposition of a temporary or permanent clinical hold by Regulatory Authorities;
•
selection of clinical trial end points that require prolonged periods of clinical observation or analysis of the resulting data;
•
clinical trials producing negative or inconclusive results;
•
a facility manufacturing our product candidates or any of their components being ordered by the FDA or comparable foreign authorities to temporarily or permanently shut down due to violations of cGMP regulations or other applicable requirements, contamination or cross-contamination of product candidates in the manufacturing process;
•
third-party clinical investigators losing the licenses or permits necessary to perform our clinical trials, not performing our clinical trials on our anticipated schedule or consistent with the clinical trial protocol or other regulatory requirements or committing fraud; or
•
changes in regulatory requirements, guidance, or feedback from Regulatory Authorities that require amending or submitting new clinical protocols or otherwise modifying the design of our clinical trials.
We could also encounter delays if a clinical trial is suspended or terminated by us, by the IRBs or ethics committees overseeing the conduct of such trials, by a Data Safety Monitoring Board for such trial or by Regulatory Authorities. Such Regulatory Authorities may impose such a suspension or termination due to a number of factors, including failure to conduct the clinical trial in accordance with regulatory requirements or our clinical protocols, inspection of the clinical trial operations or trial site by Regulatory Authorities resulting in the imposition of a clinical hold, unforeseen safety issues or adverse side effects, failure to demonstrate a benefit from using a drug, changes in governmental regulations or administrative actions or lack of adequate funding to continue the clinical trial. In addition, changes in regulatory requirements and policies may occur, and we may need to amend clinical trial protocols to comply with these changes. Amendments may require us to resubmit our clinical trial protocols to IRBs or ethics committees for reexamination and approval, which may impact the costs, timing or successful completion of a clinical trial.
Further, conducting clinical trials in foreign countries, as we are doing and expect to continue to do for our product candidates, presents additional risks that may delay completion of our clinical trials. These risks include the failure of enrolled patients in foreign countries to adhere to clinical protocols as a result of differences in healthcare services or cultural customs, managing additional administrative burdens associated with foreign regulatory requirements, as well as political, currency exchange and other economic risks relevant to such foreign countries. We may face delays in meeting our anticipated timelines for our ongoing and planned clinical trials, which could adversely affect our business, financial condition, results of operations and prospects.
Any inability to successfully complete preclinical and clinical development could result in additional costs to us or impair our ability to generate revenue from future product sales and regulatory and commercialization milestones. In addition, if we make manufacturing or formulation changes to our product candidates, we may need to conduct additional testing to bridge our modified product candidate to earlier versions. Clinical trial delays could also shorten any periods during which we may have the exclusive right to commercialize our product candidates, if approved, or allow our competitors to bring comparable products to market before we do, which could impair our ability to successfully commercialize our product candidates and may harm our business, financial condition, results of operations and prospects.
We may expend our limited resources to pursue a particular product candidate and fail to capitalize on product candidates or indications that may be more profitable or for which there is a greater likelihood of success.
We have limited financial and managerial resources, and to date, we have focused our research and development efforts on RNAi therapeutics targeting thromboembolic disease, Stargardt disease,
and anemia of chronic disease. Correctly prioritizing our research and development activities is particularly important for us due to the breadth of potential product candidates and indications that we believe could be pursued by leveraging our City Platform. As a result, we may forgo or delay pursuit of opportunities with other product candidates or in other therapeutic areas that later prove to have greater commercial potential. Our resource allocation decisions may cause us to fail to timely capitalize on viable commercial products or profitable market opportunities. Our spending on current and future research and development programs and product candidates for specific indications may not yield any commercially viable products. If we do not accurately evaluate the commercial potential or target market for a particular product candidate, we may relinquish valuable rights to that product candidate through collaboration, licensing or other royalty arrangements in cases in which it would have been more advantageous for us to retain sole development and commercialization rights to such product candidate. We must continually assess the potential commercial viability of our research programs and product candidates, and we may decide to pause or discontinue development of any of our product candidates based upon such assessments, even if we obtain positive data from our product candidates in preclinical studies and clinical trials. Similarly, our decisions to delay, terminate, or collaborate with third parties in respect of certain drug development programs may also prove not to be optimal and could cause us to miss valuable opportunities.
We are dependent on the success of our product candidates, including CITY-FXI, CITY-RBP4 and CITY-TFR2, and our ongoing and anticipated clinical trials may not be successful.
Our future success is dependent on our ability to successfully develop and timely obtain marketing approval for, and then successfully commercialize, our current product candidates, including CITY-FXI, CITY-RBP4 and CITY-TFR2. We are investing a substantial portion of our financial and managerial resources in the development of CITY-FXI, our lead product candidate, as well as CITY-RBP4 and CITY-TFR2, including our ongoing Phase 1 clinical trial for CITY-FXI, our planned Phase 1 clinical trial initiation for CITY-RBP4 in late 2026, our planned IIT initiation for CITY-TFR2 in China in late 2026 and our planned IND or IND-equivalent application submission for CITY-TFR2 in late 2026 and Phase 1/2 clinical trial initiation in early 2027.
Our product candidates will require additional clinical development, evaluation of clinical, preclinical and manufacturing activities, marketing approval in multiple jurisdictions, substantial investment and significant marketing efforts before we generate any revenues from product sales. We are not permitted to market or promote these product candidates, or any other product candidates, before we receive marketing approval from the applicable Regulatory Authorities, and we may never receive such marketing approvals.
The success of our product candidates will depend on a variety of factors. We do not have control over many of these factors, including certain aspects of clinical development and the regulatory submission and review process, potential threats to our intellectual property rights and our manufacturing, marketing, distribution and sales efforts or those of any current or future collaborator. In addition, we do not have control over whether products that target the same indications as our product candidates are introduced, which could impact the competitiveness of our product candidates. Accordingly, we cannot assure you that we will ever be able to generate revenue through the sale of these product candidates, even if approved. If we are not successful in developing and commercializing CITY-FXI, CITY-RBP4 or CITY-TFR2, or any other product candidate, or are significantly delayed in doing so, our business will be materially harmed.
Our current and future clinical trials or those of our future collaborators may reveal significant AEs or undesirable side effects not seen in our preclinical studies and may result in a safety profile that could halt clinical development, inhibit regulatory approval, limit commercial potential, or market acceptance of any of our current or future product candidates.
There is typically an extremely high rate of attrition for product candidates across categories of medicines proceeding through clinical trials. These product candidates may fail to show the desired safety and efficacy or safety, purity and potency profile in later stages of clinical trials despite having progressed through preclinical studies and initial clinical trials. A number of companies in the
biopharmaceutical industry have suffered significant setbacks in later-stage clinical trials due to lack of efficacy or unacceptable safety profiles, notwithstanding promising results in earlier trials. Most product candidates that commence clinical trials are never approved as products and there can be no assurance that any of our current or future clinical trials will ultimately be successful or support further clinical development of any of our product candidates.
Undesirable side effects caused by our product candidates, whether used alone or in combination with other therapies, could cause us or Regulatory Authorities to interrupt, delay or halt clinical trials and could result in a more restrictive label, if approved, or otherwise result in the delay or denial of regulatory approval by Regulatory Authorities. We may observe unexpected and undesirable safety or tolerability issues with our product candidates in ongoing or future clinical trials. For example, our most advanced product candidate, CITY-FXI, is being developed to be injected subcutaneously. There are risks inherent in subcutaneous injections, such as injection-site reactions (such as redness, itching, swelling, pain and tenderness) and other side effects.
If significant AEs or unacceptable side effects are observed in any of our current or future clinical trials, we may have difficulty recruiting potential trial participants to any of our clinical trials, existing trial participants may withdraw from trials, or we may be required to abandon the trials or our development efforts of one or more product candidates altogether. We, Regulatory Authorities or an IRB or ethics committee, may impose a clinical hold, suspend or terminate clinical trials of a product candidate at any time for various reasons, including a belief that participants in such trials are being exposed to unacceptable health risks or adverse side effects. Some potential therapeutics developed in the biotechnology industry that initially showed therapeutic promise in early-stage trials have later been found to cause side effects that prevented their further development. In addition, these side effects may not be appropriately recognized or managed by the treating medical staff. We may need to train medical personnel using our product candidates to understand the side effect profiles for our clinical trials and upon any commercialization of any of our product candidates. Inadequate training in recognizing or managing the potential side effects of any of our product candidates could result in harm to patients that are administered any of our product candidates. Even if the side effects do not preclude the drug from obtaining or maintaining marketing approval, unfavorable benefit risk ratio may inhibit market acceptance of the approved product due to its tolerability versus other therapies. In addition, an extended half-life could prolong the duration of undesirable side effects, which could also inhibit market acceptance. Any of these developments could materially harm our business, financial condition, results of operations and prospects.
Moreover, clinical trials are conducted in carefully defined sets of patients who have agreed to enter into clinical trials. Consequently, it is possible that our clinical trials may indicate an apparent positive effect of a product candidate that is greater than the actual positive effect, if any, or alternatively fail to identify undesirable side effects.
In addition, even if we successfully advance our product candidates or any future product candidates through clinical trials, such trials will only include a limited number of patients and limited duration of exposure to our product candidates. As a result, we cannot be assured that adverse effects of our product candidates will not be uncovered when a significantly larger number of patients are exposed to the product candidate after approval. Further, any clinical trials may not be sufficient to determine the effect and safety consequences of using our product candidates over a multi-year period.
If any of the foregoing events occur or if one or more of our product candidates prove to be unsafe, our entire pipeline or our City Platform could be affected, which would have a material adverse effect on our business, financial condition, results of operations and prospects.
If our clinical trials fail to replicate positive results from earlier preclinical studies or clinical trials conducted by us or third parties, we may be unable to successfully develop, obtain regulatory approval for or commercialize our product candidates.
The results observed from preclinical studies or early-stage clinical trials of CITY-FXI or any other product candidates may not necessarily be predictive of the results of later-stage clinical trials that we conduct. Similarly, positive results from preclinical studies or early-stage clinical trials may not be
replicated in our subsequent preclinical studies or clinical trials. For instance, the generally well-tolerated profile and FXI knockdown observed in the initial first three cohorts of our ongoing Phase 1 clinical trial for CITY-FXI in healthy volunteers, together with the preclinical results observed in NHPs and cell-based assays, may not translate to similar results in later-stage clinical trials or in patients with thrombotic disease. In addition, the degree or duration of FXI knockdown, the tolerability profile, or any potential therapeutic benefit observed in healthy volunteers may not be replicated in larger or more diverse patient populations, and future clinical trials may identify AEs or other findings that were not observed in our early-stage studies. Even if CITY-FXI demonstrates favorable tolerability and pharmacodynamic activity in our ongoing Phase 1 study, such results may not be sufficient to support advancement into later-stage clinical development or ultimately support regulatory approval. Furthermore, our product candidates may not be able to demonstrate similar activity or AE profiles as those observed in earlier studies and trials, and we may not have generated sufficient safety data to support a marketing application by the time of our targeted submission as other third-party products or product candidates that we believe may have similar profiles. In addition, in our planned future clinical trials, we may utilize clinical trial designs or dosing regimens that have not been tested in prior clinical trials.
There can be no assurance that any of our clinical trials will ultimately be successful or support further clinical development of CITY-FXI or any other product candidates. There is a high failure rate for drugs proceeding through clinical trials. Many companies in the pharmaceutical and biotechnology industries have suffered significant setbacks in late-stage clinical trials after achieving positive results in early-stage development, and we cannot be certain that we will not face similar setbacks. These setbacks have been caused by, among other things, adverse safety or efficacy observations made in clinical trials.
Additionally, we intend to utilize an “open-label” clinical trial design for certain of our clinical trials. An “open-label” clinical trial is one where both the patient and investigator know whether the patient is receiving the investigational product candidate or either an existing approved drug or placebo. Most open-label clinical trials test only the investigational product candidate and sometimes may do so at different dose levels. Open-label clinical trials are subject to various limitations that may exaggerate any therapeutic effect as patients in open-label clinical trials are aware when they are receiving treatment. Open-label clinical trials may be subject to a “patient bias” where patients perceive their symptoms to have improved merely due to their awareness of receiving an experimental treatment. In addition, open-label clinical trials may be subject to an “investigator bias” where those assessing and reviewing the physiological outcomes of the clinical trials are aware of which patients have received treatment and may interpret the information of the treated group more favorably given this knowledge. The results from an open-label trial may not be predictive of future clinical trial results of a product candidate when studied in a controlled environment with a placebo or active control.
Moreover, preclinical and clinical data are often susceptible to varying interpretations and analyses and many companies that believed their product candidates performed satisfactorily in preclinical studies and clinical trials nonetheless failed to obtain approval from Regulatory Authorities.
Interim, initial, topline and preliminary data from our clinical trials that we announce or publish from time to time may change as more patient data become available and are subject to audit and verification procedures that could result in material changes in the final data.
From time to time, we may publicly disclose interim, preliminary or topline data from our preclinical studies and clinical trials, which are based on a preliminary analysis of then-available data, and the results and related findings and conclusions are subject to change following a more comprehensive review of the data related to the particular study or trial. We also make assumptions, estimations, calculations and conclusions as part of our analyses of data, and we may not have received or had the opportunity to fully and carefully evaluate all data. As a result, the topline or preliminary results that we report may differ from future results of the same studies, or different conclusions or considerations may qualify such results, once additional data have been received and fully evaluated. Topline and preliminary data also remain subject to audit and verification procedures that may result in the final data being materially different from the topline or preliminary data we previously made public. As a result, topline and preliminary data should be viewed with caution until the final data are available. From time to time, we
may also disclose interim data from our clinical trials. Interim data from clinical trials that we may complete are subject to the risk that one or more of the clinical outcomes may materially change as patient enrollment continues and more patient data become available. Adverse differences between topline, preliminary or interim data and final data could significantly harm our business prospects, financial condition, results of operations and prospects.
Further, others, including Regulatory Authorities, may not accept or agree with our assumptions, estimates, calculations, conclusions or analyses or may interpret or weigh the importance of data differently, which could impact the value of the particular program, the approvability or commercialization of the particular product candidate or product and our company in general. In addition, the information we choose to publicly disclose regarding a particular study or clinical trial is based on what is typically extensive information, and you or others may not agree with what we determine is the material or otherwise appropriate information to include in our disclosure, and any information we determine not to disclose may ultimately be deemed significant with respect to future decisions, conclusions, views, activities or otherwise regarding a particular product, product candidate or our business. If the topline or preliminary data that we report differ from actual results, or if others, including Regulatory Authorities, disagree with the conclusions reached, our ability to obtain approval for, and commercialize, our product candidates may be harmed, which could harm our business, operating results, prospects or financial condition. Further, disclosure of interim, topline or preliminary data by us or by our competitors could result in volatility in the price of our common stock.
The manufacture of biopharmaceutical products is a complex process which requires significant expertise and capital investment, and if we encounter problems in our collaboration with third parties for the manufacture for clinical or commercial scale or in the manufacture of our future product candidates, our business could suffer.
We currently do not have cGMP manufacturing capabilities and we are entirely dependent on third-party contractors, including contract development and manufacturing organizations (“CDMOs”), to manufacture our product candidates for our clinical trials and preclinical studies, and we expect to rely on third-party CDMOs for the production of commercial quantities of our products in accordance with cGMP regulations if we obtain regulatory and marketing approvals. The manufacture of biopharmaceutical products is a complex process, in part due to strict regulatory requirements. If we are unable to identify and maintain collaboration with third-party CDMOs, or fail to do so in a timely manner or on advantageous terms, this may lead to significant delays in the clinical supply of our product candidates as well as the commercial manufacturing of our products once regulatory and marketing approvals have been obtained, if ever. In turn, this could delay our clinical trials, negatively impact our ability to ultimately obtain regulatory approval and materially harm any future commercialization plans. Additionally, we may not be able to establish our chemistry, manufacturing and controls (“CMC”) and quality control capabilities, which are essential for regulatory compliance and product quality assurance, and any failure to develop these capabilities, including any failure to recruit and retain and motivate qualified CMC and quality control personnel, could further delay our manufacturing and commercialization efforts.
In addition, since we have no history of commercializing pharmaceutical products, we may face additional challenges in developing qualified teams and acquiring the necessary expertise to oversee third-party manufacturing processes. Any delay or failure to establish adequate CMC, quality control and manufacturing oversight capabilities may result in problems that arise during the manufacturing process for a variety of reasons, including, but not limited to, equipment malfunction, failure to follow specific protocols and procedures, problems with (including shortage of) raw materials, global supply chain issues (such as trade barriers, export restrictions and other impediments to the cross-border movement of materials and goods), and limits to manufacturing capacity due to regulatory requirements, changes in the types of products produced, increases in the prices of raw materials, physical limitations that could inhibit continuous supply, man-made or natural disasters and environmental factors. If problems arise during the production of a batch of future products, that batch of future products may have to be discarded and we may experience product shortages or incur added expenses. This, as well as problems that may arise during the manufacturing process, could, among other things, lead to significant additional costs and/or delays, lost revenue, damage to customer relationships, time and
expense spent investigating the cause and, depending on the cause, similar losses with respect to other batches or product candidates. If problems are not discovered before such product is released to the market, recall and product liability costs may also be incurred.
Our future growth may depend, in part, on our ability to operate in foreign markets, where we would be subject to additional regulatory burdens and other risks and uncertainties.
Our future growth may depend, in part, on our ability to develop and commercialize our product candidates in foreign markets for which we may rely on collaboration with third parties. Recent and ongoing changes in the United States trade policy with foreign countries, including the continued uncertainty surrounding United States tariffs and potential retaliatory measures by foreign governments may disrupt the global supply chain for biopharmaceutical products. For example, in September 2025, President Trump announced plans to impose 100% tariffs on imported branded or patented pharmaceuticals, unless the importing company is building United States manufacturing capacity. It is not yet clear whether these tariffs would apply to the importation of active pharmaceutical ingredients (“APIs”) and possibly bulk drug products that are intended for use in clinical trials and not for commercial sale, which could increase the costs of materials for our clinical trials. Any direct tariffs, if imposed on pharmaceutical products, may result in increased costs for raw materials and contract manufacturing services, reduced ability to source critical CDMO arrangements, and a delay in our development timelines.
We are not permitted to market or promote any of our product candidates before we receive regulatory approval from the applicable foreign regulatory authority and may never receive such regulatory approval for any of our product candidates. To obtain separate regulatory approval in many other countries, we must comply with numerous and varying regulatory requirements of such countries regarding safety and efficacy and governing, among other things, clinical trials and commercial sales, pricing and distribution of our product candidates, and we cannot predict success in these jurisdictions. If we fail to comply with the regulatory requirements in international markets and receive applicable marketing approvals, our target market will be reduced and our ability to realize the full market potential of our product candidates will be harmed and our business will be adversely affected. Moreover, even if we obtain approval of our product candidates and ultimately commercialize our product candidates in foreign markets, we would be subject to the risks and uncertainties, including the burden of complying with complex and changing foreign regulatory, tax, accounting and legal requirements and reduced protection of intellectual property rights in some foreign countries.
The biopharmaceutical market is intensely competitive. If we or our collaborators are unable to compete effectively with existing drugs, new treatment methods, and new technologies, including with respect to our specific product candidates CITY-FXI, CITY-RBP4 and CITY-TFR2, we may be unable to successfully commercialize any drugs that we develop.
The biopharmaceutical industry is characterized by rapidly advancing technologies, intense competition, and a strong emphasis on proprietary and novel products and product candidates. We face a broad spectrum of current and potential competitors, ranging from very large, global pharmaceutical companies with significant resources to other biotechnology companies with resources and expertise comparable to our own. We believe that for most or all of our drug development programs, there will be one or more competing programs in other companies. In many cases, the companies with competing programs will have access to greater resources and expertise than we do and may be more advanced. As RNAi therapies expand beyond rare diseases to other more common conditions, an increasing number of companies are investing in and developing RNAi therapeutics, as well as potentially new approaches that may result in more rapid development of RNAi therapeutics or more effective technologies for RNAi drug development or delivery. Companies that have developed, or are developing, siRNA therapeutics include ADARx Pharmaceuticals, Inc., Alnylam, Amgen Inc., Argo Biopharmaceutical Co., Ltd., Aro Biotherapeutics Co., Arrowhead Pharmaceuticals, Inc., AstraZeneca plc, Dyne Therapeutics, Inc., Eli Lilly and Company, GlaxoSmithKline plc, Ionis Pharmaceuticals, Inc., Novartis AG, Novo Nordisk A/S, SanegeneBio Inc., Sarepta Therapeutics, Inc., Silence Therapeutics plc, and Sirius Therapeutics, Inc. Many of our competitors have:
•
substantially greater financial, technical and human resources than we have;
•
more extensive experience in preclinical testing, conducting clinical trials, obtaining regulatory approvals and in manufacturing, marketing and selling products;
•
product candidates that are based on previously tested or accepted technologies;
•
products that have been approved or are in late stages of development; and
•
collaborative arrangements in our target markets with leading companies and research institutions.
Accordingly, our competitors may be more successful than us in obtaining patent protection, regulatory exclusivities or FDA approval and may commercialize products or achieve widespread market acceptance more rapidly than we do, which may impact future approvals or sales of our product candidates that receive regulatory approval. If the FDA approves the commercial sale of our product candidates, we will also be competing with respect to marketing capabilities and manufacturing efficiency. We expect competition among products will be based on product efficacy and safety, the timing and scope of regulatory approvals, availability of supply, marketing and sales capabilities, product price, reimbursement coverage by government and private third-party payors, regulatory exclusivities and patent position, among other factors. Our profitability and financial position will suffer if our product candidates receive regulatory approval but cannot compete effectively in the marketplace. Our competitors have developed, are developing or may develop product candidates and products competitive with CITY-FXI, CITY-RBP4 and CITY-TFR2 and our other programs and product candidates. CITY-FXI, CITY-RBP4 and CITY-TFR2 and any future product candidates that we successfully develop and commercialize will compete with existing therapies and new therapies that may become available in the future.
Our lead product candidate, CITY-FXI, is being developed for thromboembolic diseases through silencing of coagulation FXI. Multiple therapeutic approaches targeting FXI are currently in development, including small molecules, monoclonal antibodies and RNAi therapeutics. Companies active in this field include ADARx Pharmaceuticals, Inc., Bayer AG, Bristol-Myers Squibb Company, CRISPR Therapeutics AG (in partnership with Sirius Therapeutics, Inc.), Johnson & Johnson, Novartis AG, Regeneron Pharmaceuticals, Inc., and others. CITY-RBP4 is being developed for Stargardt disease, for which current treatment options are limited. Numerous companies are developing potential therapies for Stargardt disease using a variety of approaches, including visual cycle modulation, gene therapy, gene editing and RNA-based therapeutics. Companies active in this area include Alkeus Pharmaceuticals, Inc., Belite Bio, Inc. and others. CITY-TFR2 is being developed for anemia of chronic disease. Therapeutic approaches under development or in clinical use include hepcidin-targeted agents, including a monoclonal antibody from Disc Medicine, Inc., erythropoiesis-stimulating agents, hypoxia-inducible factor prolyl hydroxylase inhibitors, iron replacement therapies and other approaches intended to improve iron availability or erythropoiesis.
These drugs may be more effective, safer, less expensive or marketed and sold more effectively, than any products we develop. In most cases, we do not currently plan to run head-to-head clinical trials evaluating our product candidates against the current standards of care, which may make it more challenging for our product candidates to compete against the current standards of care due to the lack of head-to-head clinical trial data.
If we successfully develop any product candidates, and obtain approval for them, we expect to face competition based on many different factors, including: the safety and effectiveness of our products relative to alternative therapies, if any; the ease with which our products can be administered and the extent to which patients accept relatively new routes of administration; the timing and scope of regulatory approvals for these product candidates; the availability and cost of manufacturing, marketing and sales capabilities; the price of any approved product; reimbursement coverage; and patent position. See the section titled “Business — Competition” included elsewhere in this prospectus for examples of the competition that we face.
In addition, our competitors may develop partnership, collaboration and licensing arrangements with or receive funding from larger pharmaceutical or biotechnology companies, providing them with an
advantage over us. Our competitors may also succeed in developing, acquiring or licensing technologies and drug products that are more effective or less costly than our product candidates, which could render our product candidates obsolete and noncompetitive. Our competitors may therefore be more successful in commercializing their products than we are, which could adversely affect our competitive position and business. Competitive products may make any products we develop obsolete or noncompetitive before we can recover the expenses of developing and commercializing our products, if approved.
Mergers and acquisitions in the pharmaceutical and biotechnology industries may result in even more resources being concentrated among a smaller number of our competitors. Smaller and other early-stage companies may also prove to be significant competitors, particularly through collaborative arrangements with large and established companies. Moreover, we may also compete with universities, governmental agencies and other public and private research institutions that may be active in research in our target indications and could be in direct competition with us. These third parties compete with us in recruiting and retaining qualified scientific, management and clinical development personnel and establishing clinical trial sites, as well as in acquiring technologies complementary to, or necessary for, our product candidates, and in identifying and in-licensing intellectual property related to new product candidates, as well as entering into partnerships, collaborations and other license arrangements. Our inability to compete successfully for such personnel, sites, intellectual property and arrangements could negatively affect our level of expertise and our ability to execute our business plan.
We and our collaborators face competition from other companies that are working to develop novel drugs and technology platforms using technology similar to ours, as well as from companies utilizing emerging technologies. If these companies develop drugs more rapidly than we or our collaborators do or their technologies, including delivery technologies, are more effective, our and our collaborators’ ability to successfully commercialize our products may be adversely affected.
In addition to competition from companies developing therapies for the same indications, we and our collaborators face competition from other companies developing RNAi therapeutics and other technologies that compete more directly with our own. As RNAi therapies expand beyond rare diseases to other more common conditions, an increasing number of companies are investing in and developing RNAi therapeutics, as well as potentially new approaches that may result in more rapid development of RNAi therapeutics or more effective technologies for RNAi drug development or delivery. We are aware of several other companies that are working to develop RNAi therapeutics. Some of these companies are seeking, as we are, to develop chemically synthesized siRNAs as drugs. Others are following a gene therapy approach, with the goal of treating patients with synthetic, exogenously-introduced genes designed to produce siRNA-like molecules within cells. Our competitors may develop RNAi technology more rapidly and more effectively than we do.
In addition to competition with respect to RNAi and specific product candidates, we face substantial competition to discover and develop safe and effective means of delivering RNAi therapeutics to the relevant cells and tissues. If our competitors develop superior or more effective delivery technologies, our ability to successfully develop and commercialize our product candidates could be adversely affected. In addition, third parties, including private companies and academic institutions, are expending substantial resources to discover and develop safe and effective means of delivering RNAi therapeutics to the relevant cells and tissues. Some of our competitors have substantially greater resources than we do, and if they obtain exclusive rights to delivery technologies developed by third parties, our ability to develop and successfully commercialize our product candidates could be adversely affected.
If the market opportunities for our product candidates are smaller than we believe they are, our revenue may be adversely affected and our business may suffer.
The estimates of market opportunity and forecasts of market growth included in documents that we file with the SEC may prove to be smaller than we believe, and even if the markets in which we compete achieve the forecasted growth, our business may not grow at similar rates, or at all. Our pipeline is
focused on developing RNAi therapeutics designed to suppress disease-relevant hepatic and extra-hepatic targets for diseases with significant unmet medical need, including thromboembolic disease, Stargardt disease, and anemia of chronic disease. Our projections of both the number of people who have these diseases, as well as the subset of people with these diseases who have the potential to benefit from treatment with our product candidates, are based on estimates and independent market research, industry and general publications obtained from third parties. Market opportunity estimates and growth forecasts included in this prospectus and the other documents that we file with the SEC are subject to significant uncertainty and are based on assumptions and estimates. These estimates, which have been derived from a variety of sources, including scientific literature, surveys of clinics, patient foundations and market research, may prove to be incorrect. Further, new studies may change the estimated incidence or prevalence of these indications. Additionally, the potentially addressable patient population may not ultimately be amenable to treatment with our product candidate if we cannot achieve our intended dosing interval. Our market opportunity may also be limited by current and future products of our competitors that are already available in the market or may enter the market for such patients. If any of our estimates prove to be inaccurate, the market opportunity for our product candidates could be significantly diminished and have an adverse material impact on our business.
The market may not be receptive to our product candidates based on a novel therapeutic modality, and we may not generate any future revenue from the sale, licensing, or royalties related to product candidates.
Even if approval is obtained for a product candidate, we may not generate or sustain revenue from sales, licensing, or royalties related to the product due to factors such as whether the product can be sold at a competitive cost and otherwise accepted in the market. The product candidates that we are developing are based on new technologies and therapeutic approaches. Market participants with significant influence over acceptance of new treatments, such as physicians and third-party payors, may not adopt a treatment based on siRNA technology for the indications we are targeting, and we may not be able to convince the medical community and third-party payors to accept and use, or to provide favorable reimbursement for, our product candidates. Market acceptance of our product candidates will depend on, among other factors: the timing of our receipt of any marketing and commercialization approvals; the terms of any approvals and the countries in which approvals are obtained; the clinical indications for which our product candidates are approved; the safety and efficacy of our product candidates, and physicians’ and patients’ perception of our product candidates as a safe and effective treatment; the frequency and severity of any adverse side effects associated with our product candidates; limitations or warnings contained in any labeling approved by Regulatory Authorities; relative convenience and ease of administration of our product candidates; the willingness of patients to accept any new methods of administration; the success of our physician education programs; the availability of adequate government and third-party payor reimbursement; the pricing of our products, particularly as compared to alternative treatments; the potential and perceived advantages of our product candidates over alternative treatments; availability of alternative effective treatments for the disease indications our product candidates are intended to treat and the relative risks, benefits and costs of those treatments; the inclusion in government-sponsored medical insurance programs, or by third-party payors; the willingness of patients to pay out-of-pocket in the absence of coverage and reimbursement by third-party payors and government authorities; and the effectiveness of our sales and marketing efforts.
With our focus on the growing field of RNAi therapeutics, these risks may increase to the extent the space becomes more competitive or less favored in the commercial marketplace. Market size is also a variable in disease indications not classified as rare. Our estimates regarding potential market size for any indication may be materially different from what we discover to exist at the time it commences commercialization, if any, for a product, which could result in significant changes in our business plan and have a material adverse effect on our business, financial condition, results of operations and prospects. Even if a potential product displays a favorable efficacy and safety profile in preclinical and clinical studies, market acceptance of the product will not be fully known until after it is launched. Our efforts to educate the medical community and third-party payors on the benefits of the product candidates may require significant resources and may never be successful. If our product candidates are approved but
fail to achieve an adequate level of acceptance by physicians, patients, third-party payors, and others in the medical community, we will not be able to generate sufficient revenue for us to become or remain profitable.
Also, we may not generate any future revenue from the sale or licensing of product candidates as we have no track record with no experience in launching and marketing approved drugs. Our financial performance depends on the successful development, regulatory approval and subsequent launching and marketing of our product candidates, which are in early-stage clinical and preclinical development. As all of our drug candidates are in the development stage, we have not yet demonstrated an ability to commercialize any of our drug candidates, if approved. Our ability to successfully commercialize approved drugs may involve more inherent risk, take longer, and cost more than it would if we were a company with experience launching and marketing approved drugs.
We may decide to build up our in-house commercialization and distribution capabilities to maximize our product offering and expedite market acceptance of our products. In such case, we will have to compete with other pharmaceutical and biopharmaceutical companies to recruit, hire, train and retain marketing and sales personnel. There can be no assurance that we will be able to develop and successfully maintain in-house sales and commercial distribution capabilities to successfully commercialize any of our product candidates, if and when approved, and as a result, we may not be able to generate sales revenue as planned.
If we are unable to, or decide not to, develop internal sales, marketing and commercial distribution capabilities, we will likely pursue collaborative arrangements regarding the sales and marketing of our approved drugs. However, there can be no assurance that we will be able to establish or maintain such collaborative arrangements, or that we will have effective sales forces after establishing such collaborative arrangements. Any revenue we receive will depend upon the efforts of such third parties. We would have little or no control over the marketing and sales efforts of such third parties, and our revenue from product sales may be lower than if we had commercialized our product candidates ourselves in a cost-effective manner. We also face competition in our search for third parties to assist us with the sales and marketing efforts for our product candidates. In case we cannot develop and successfully maintain in-house sales and commercial distribution capabilities or collaborate with third parties to successfully commercialize our products, we may not be able to generate product sales revenue and our business and prospects may suffer.
The insurance coverage and reimbursement status of newly approved products, in a new category of medicines, is uncertain. Failure to obtain or maintain adequate coverage and reimbursement for new or current products could limit our ability to market those products and decrease our ability to generate revenue.
The availability and extent of reimbursement by governmental and private payors is essential for most patients to be able to afford expensive treatments such as the products that we hope to develop and sell. Sales of our product candidates, if approved and commercialized, will depend substantially, both domestically and abroad, on the extent to which the costs of our product candidates will be paid by health maintenance, managed care, pharmacy benefit and similar healthcare management organizations or reimbursed by government health administration authorities, private health coverage insurers and other third-party payors. Even if coverage is provided, the approved reimbursement amount may not be high enough to allow us to establish or maintain pricing sufficient to realize a sufficient return on our investment in any of our products. If reimbursement is not available, or is available only to limited levels, we may not be able to successfully commercialize our product candidates.
There is significant uncertainty related to the insurance coverage and reimbursement of newly approved products. Government authorities and other third-party payors, such as private health insurers and health maintenance organizations, decide which drugs and treatments they will cover and the amount of reimbursement. Coverage and reimbursement by a third-party payor may depend upon a number of factors, including the third-party payor’s determination that use of a product is: a covered benefit under its health plan; safe, effective and medically necessary; appropriate for the specific patient; cost-effective; and neither experimental nor investigational.
In the United States, no uniform policy of coverage and reimbursement for products exists among third-party payors. The Centers for Medicare & Medicaid Services (“CMS”), an agency within the United States Department of Health and Human Services (“HHS”), determines whether and to what extent a new medicine will be covered and reimbursed under Medicare. Private payors tend to follow CMS to a substantial degree. It is difficult to predict what CMS will decide with respect to reimbursement for products.
Net prices for drugs may be reduced by mandatory discounts or rebates required by government healthcare programs or private payors and by any future relaxation of laws that presently restrict imports of drugs from countries where they may be sold at lower prices than in the United States. Increasingly, third-party payors are requiring that drug companies provide them with predetermined discounts from list prices and are challenging the prices charged for medical products. We cannot be sure that reimbursement will be available for our product candidates that we commercialize and, if reimbursement is available, the level of reimbursement. In addition, many biopharmaceutical manufacturers must calculate and report certain price reporting metrics to the government, such as average sales price and best price. Penalties may apply in some cases when such metrics are not submitted accurately and timely. Further, these prices for drugs may be reduced by mandatory discounts or rebates required by government healthcare programs. Payment methodologies may be subject to changes in healthcare legislation and regulatory initiatives.
Outside the United States, certain countries, including a number of EU Member States, set prices and reimbursement for pharmaceutical products, or medicinal products, as they are commonly referred to in the European Union, with limited participation from the marketing authorization holders. Reimbursement agencies in Europe may be more conservative than CMS. For example, a number of cancer drugs have been approved for reimbursement in the United States and have not been approved for reimbursement in certain European countries. We cannot be sure that such prices and reimbursement will be acceptable to us or our collaboration partners. If Regulatory Authorities in these foreign jurisdictions set prices or reimbursement levels that are not commercially attractive for us or our collaboration partners, our revenues from sales by us or our collaboration partners and the potential profitability of our products in those countries would be negatively affected. An increasing number of countries are taking initiatives to attempt to reduce large budget deficits by focusing cost-cutting efforts on pharmaceuticals for their state-run health care systems. These international price control efforts have impacted all regions of the world but have been most drastic in the European Union.
Additionally, the requirements governing product pricing vary widely from country to country. Some countries require approval of the sale price of a product before it can be marketed, while in others, the pricing review period begins after marketing or product licensing approval is granted. As a result, we might obtain marketing approval for a product in a particular country, but then may experience delays in the reimbursement approval of our product or be subject to price regulations that would delay our commercial launch of the product, possibly for lengthy time periods, which could negatively impact the revenues we are able to generate from the sale of the product in that particular country. For example, the European Union provides options for its Member States to restrict the range of medicinal products for which their national health insurance systems provide reimbursement and to control the prices of medicinal products for human use. To obtain reimbursement or pricing approval, some of these countries may require the completion of clinical trials that compare the cost effectiveness of a particular product candidate to currently available therapies. A Member State may approve a specific price for the medicinal product or it may instead adopt a system of direct or indirect controls on the profitability of the company placing the medicinal product on the market. There can be no assurance that any country that has price controls or reimbursement limitations for pharmaceutical products will allow favorable reimbursement and pricing arrangements for our product candidates. Historically, products launched in the European Union do not follow price structures of the United States and generally prices tend to be significantly lower.
Moreover, increasing efforts by governmental and third-party payors, in the United States and abroad, to cap or reduce healthcare costs may cause such organizations to limit both coverage and level of reimbursement for new products approved and, as a result, they may not cover or provide adequate payment for our product candidates.
We expect to experience pricing pressures in connection with the sale of any of our product candidates, if approved and commercialized, due to the trend toward managed healthcare, the increasing
influence of health maintenance organizations and additional legislative changes. The downward pressure on healthcare costs in general, particularly prescription drugs and surgical procedures and other treatments, has become very intense. As a result, increasingly high barriers are being erected to the entry of new products. The implementation of cost containment measures or other healthcare reforms may prevent us from being able to generate revenue, attain profitability or commercialize our products.
For more information on the laws and regulations that may impact coverage and reimbursement of our product candidates, see the section titled “Business — Government Regulation — Coverage and Reimbursement” and “— Healthcare Reform and Other Regulatory Changes” included elsewhere in the prospectus.
Our operations, including our relationships with healthcare providers, physicians and third-party payers, are subject to applicable anti-kickback, fraud and abuse, and other healthcare laws and regulations, which, in the event of a violation, exposes us to liability for criminal sanctions, civil penalties, and contractual damages, and reputational harm and diminished profits and future earnings.
Our operations, including any arrangements that we enter into with healthcare providers, physicians, and third-party payers, are subject to broadly applicable fraud and abuse and other healthcare laws and regulations. Such laws and regulations, including applicable United States federal and state healthcare laws and regulations, as well as foreign laws, such as the federal Anti-Kickback Statute, the False Claims Act, the Health Insurance Portability and Accountability Act of 1996 (“HIPAA”), the Physician Payments Sunshine Act, or the Foreign Corrupt Practices Act, may constrain our operation and the business or financial arrangements through which we can market, sell and distribute any product candidates for which we obtain marketing approval. For more information, please see “Business — Government Regulation and Product Approval-Other Healthcare Laws and Compliance Requirements.”
Efforts to confirm that our business arrangements with third parties comply with applicable healthcare laws and regulations involve substantial costs. It is possible that governmental authorities will conclude that our business practices may not comply with current or future statutes, regulations or case law involving applicable fraud and abuse or other healthcare laws and regulations. If our operations are found to be in violation of any of these laws or any other governmental regulations that may apply to us, we may become subject to significant civil, criminal and administrative penalties, damages, fines, imprisonment, exclusion of products from government funded healthcare programs, such as Medicare and Medicaid, and the curtailment or restructuring of our operations, and compliance and reporting obligations that could adversely affect our revenues, financial condition or results of operations. If any of the physicians or other healthcare providers or entities with whom we expect to do business are found to be not in compliance with applicable laws, they may be subject to criminal, civil or administrative sanctions, including exclusions from government funded healthcare programs.
If we fail to comply with environmental, health and safety laws and regulations, we could become subject to fines or penalties or incur costs that could harm our business.
We and our current and future third-party CDMOs are subject to numerous environmental, health and safety laws and regulations, including those governing laboratory procedures and the handling, use, storage, treatment and disposal of hazardous materials and wastes. Our operations will involve the use of hazardous and flammable materials, including chemicals and biological materials. Our operations also may produce hazardous waste products. We generally anticipate contracting with third parties for the disposal of these materials and wastes. We will not be able to eliminate the risk of contamination or injury from these materials. In the event of contamination or injury resulting from any use by us of hazardous materials, we could be held liable for any resulting damages, and any liability could exceed our resources. We also could incur significant costs associated with civil or criminal fines and penalties for failure to comply with such laws and regulations.
Although we maintain general liability insurance as well as workers’ compensation insurance to cover us for costs and expenses we may incur due to injuries to our employees resulting from the use of hazardous materials, this insurance may not provide adequate coverage against potential liabilities.
We do not maintain insurance for environmental liability or toxic tort allegations that may be asserted against us in connection with our storage or disposal of biological or hazardous materials.
In addition, we may incur substantial costs in order to comply with current or future environmental, health and safety laws and regulations. These current or future laws and regulations may impair our research, development or production efforts. Our failure to comply with these laws and regulations also may result in substantial fines, penalties or other sanctions.
Further, with respect to the operations of our current and any future CDMOs, it is possible that if they fail to operate in compliance with applicable environmental, health and safety laws and regulations or properly dispose of wastes associated with our products, we could be held liable for any resulting damages, suffer reputational harm or experience a disruption in the manufacture and supply of our product candidates. In addition, our supply chain may be adversely impacted if any of our CDMOs become subject to injunctions or other sanctions as a result of their non-compliance with environmental, health and safety laws and regulations.
Risks Related to Government Regulatory and Legal Requirements
While we may in the future seek designations for our product candidates with Regulatory Authorities that are intended to confer benefits such as a faster development process, a streamlined review or regulatory exclusivity, there can be no assurance that we will successfully obtain such designations. In addition, even if our product candidates are granted such designations, we may not be able to realize the intended benefits of such designations, or maintain such designations.
Regulatory Authorities offer certain designations for product candidates that are designed to encourage the research and development of product candidates that are intended to address serious conditions. These designations may confer benefits such as additional interaction with Regulatory Authorities, streamlined development pathways and expedited review procedures. However, there can be no assurance that we will qualify for or successfully obtain such designations for our product candidates. In addition, while such designations could expedite the development or approval process, they generally do not change the standards for approval. Even if we obtain such designations for our product candidates, there can be no assurance that we will realize their intended benefits.
We may also seek Breakthrough Therapy Designation for any product candidate that we develop. A breakthrough therapy is defined as a drug that is intended, alone or in combination with one or more other drugs, to treat a serious or life-threatening disease or condition, and preliminary clinical evidence indicates that the drug may demonstrate substantial improvement over currently approved therapies on one or more clinically significant endpoints, such as substantial treatment effects observed early in clinical development. For drugs that have been designated as breakthrough therapies, interaction and communication between the FDA and the sponsor of the trial can help to identify the most efficient path for clinical development while minimizing the number of patients placed in ineffective control regimens. Drugs designated as breakthrough therapies by the FDA are also eligible for accelerated approval and priority review.
Designation as a breakthrough therapy is within the discretion of the FDA. Accordingly, even if we believe a product candidate we develop meets the criteria for designation as a breakthrough therapy, the FDA may disagree and instead determine not to make such designation. In any event, the receipt of Breakthrough Therapy Designation for a product candidate may not result in a faster development process, review, or approval compared to drugs considered for approval under conventional FDA procedures and does not assure ultimate approval by the FDA. In addition, even if any product candidate we develop qualifies as a breakthrough therapy, the FDA may later decide that the drug no longer meets the conditions for qualification and rescind the designation.
Even in the absence of obtaining certain designations, a sponsor can seek priority review at the time of submitting a marketing application. The FDA may designate an application for priority review if the product is intended to treat a serious condition and, if approved, would provide a significant improvement in safety or effectiveness when compared with other available therapies. Significant
improvement may be illustrated by evidence of increased effectiveness in the treatment of a condition, elimination or substantial reduction of a treatment-limiting adverse reaction, documented enhancement of patient compliance that may lead to improvement in serious outcomes, or evidence of safety and effectiveness in a new subpopulation. A priority review designation is intended to direct overall attention and resources to the evaluation of such applications, and to shorten the FDA’s goal for acting on a marketing application from ten months to six months. Priority review designation may be rescinded if a product no longer meets the qualifying criteria.
We may in the future seek orphan drug designation for our product candidates. However, even if we obtain orphan drug designation, we may be unable to maintain the associated benefits, including the potential for market exclusivity.
We may seek orphan drug designation or exclusivity in certain indications targeted by our current or future product candidates. Regulatory authorities in some jurisdictions, including the United States and Europe, may designate drugs for relatively small patient populations as orphan drugs. For example, under the Orphan Drug Act, the FDA may designate a product candidate as an orphan drug if it is intended to treat a rare disease or condition. In order for the FDA to grant orphan drug exclusivity to one of our product candidates, the agency must find that the product candidate is indicated for the treatment of a condition or disease that affects fewer than 200,000 individuals in the U.S. or that affects 200,000 or more individuals in the U.S. and for which there is no reasonable expectation that the cost of developing and making the product candidate available for the disease or condition will be recovered from sales of the product in the U.S. Orphan drug designation must be requested before submitting an NDA. The FDA may conclude that the condition or disease for which we seek orphan drug exclusivity does not meet the required standard.
If a product that has orphan drug designation subsequently receives the first FDA approval for a particular drug for the disease for which it has such designation, the product is entitled to orphan product exclusivity, which means that the FDA may not approve any other applications to market the same drug for the same approved use or indication for seven years, except in limited circumstances, such as if the FDA finds that the holder of the orphan drug exclusivity has not shown that it can assure the availability of sufficient quantities of the orphan drug to meet the needs of patients with the disease or condition for which the drug was designated.
In addition, even after an orphan drug is approved, the FDA can subsequently approve the same product candidate for the same condition if the FDA concludes that the later product candidate is clinically superior in that it is shown to be safer, more effective or makes a major contribution to patient care compared with the product that has orphan exclusivity. In the U.S., orphan drug designation entitles a party to financial incentives such as opportunities for grant funding towards clinical trial costs, tax advantages, and user-fee waivers. After the FDA grants orphan drug designation, the generic identity of the drug or biologic and its potential orphan use are disclosed publicly by the FDA. Orphan drug designation does not convey any advantage in, or shorten the duration of, the regulatory review and approval process.
A similar regulatory scheme governs approval of orphan product candidates by the EMA in the European Union. Generally, if a product with an orphan drug designation subsequently receives the first marketing approval for the indication for which it has such designation, the product is entitled to a period of marketing exclusivity, which precludes the FDA or the EMA (as applicable) from approving another marketing application for the same or another similar product candidate for the same orphan therapeutic indication for that time period. The applicable period is seven years in the U.S. and ten years in the European Union. The exclusivity period in the European Union can be reduced to six years if at the end of the fifth year it is determined that a product no longer meets the criteria for orphan designation, including if the product is sufficiently profitable so that market exclusivity is no longer justified.
We may encounter risks associated with our planned investigator-initiated trial in China for CITY-TFR2, which could delay or adversely affect the development of CITY-TFR2.
Our development strategy for CITY-TFR2 includes a planned IIT in China to establish initial clinical proof-of-concept in patients with anemia associated with myelofibrosis. IITs are clinical studies that are
initiated, sponsored and conducted by independent investigators and/or institutions rather than by us, with the applicable investigators and/or institutions generally responsible for the conduct of the study and compliance with applicable requirements. IITs are governed under the oversight of the National Health Commission in China and implemented within accredited medical institutions. Approval by the local ethics committee and compliance with China Good Clinical Practice are mandatory requirements for IIT conduct. We expect the IIT to complement our planned IND or IND-equivalent application submission and Phase 1/2 clinical trial in other geographies. Because the IIT is an important component of our early clinical development strategy for CITY-TFR2, delays, operational challenges, regulatory issues, or an inability to obtain or utilize data from the IIT could delay the development of CITY-TFR2, affect our ability to advance the program into later-stage clinical development, or otherwise adversely affect our business.
In July 2026, we entered into an IIT study agreement with Overland Therapeutics (SH) Co. Ltd. (“Overland”), a biotechnology company that has operations in the United States and China, to support the conduct of the IIT. Pursuant to the agreement, we will provide support for the IIT, supply CITY-TFR2 and fund covered study-related services and expenses through milestone payments. Overland, as local study sponsor, will provide clinical and operational support in connection with the IIT, including supporting site and investigator selection, administrative submissions, project management, insurance and clinical monitoring and, subject to our prior approval, will enter into separate agreements with the applicable investigators, hospitals and research institutions in China. To facilitate the services in connection with the IIT, we are required to share certain confidential information about CITY-TFR2 with Overland, investigator(s), and/or institution(s) based in China. Although we require confidentiality obligations in our agreements with Overland, investigator(s), and/or institution(s) and take reasonable measures to protect our confidential information, we may have difficulty enforcing our rights in the event of any breach of confidentiality, or infringement or misappropriation of our confidential information. If we are unable to adequately protect our confidential information from unauthorized disclosure, infringement, or misappropriation, our development and intellectual property protection of CITY-TFR2 could be adversely affected. The IIT will be conducted by investigators at participating medical institutions, with clinical and operational support and coordination provided by Overland. While this IIT is not one of our traditional company-sponsored clinical trials, subject to applicable law and the applicable agreements with the investigators and medical institutions, data and results generated from the IIT will be provided to us. Although data from the IIT may support our development program for CITY-TFR2, including by providing initial clinical proof-of-concept data and informing our planned Phase 1/2 clinical trial, there can be no assurance that the FDA or other Regulatory Authorities will accept such data to support future regulatory submissions or marketing approval. We expect to report data from the IIT in mid-2027.
Given the nature of IITs, notwithstanding the support we and Overland will provide in connection with the IIT, we may have less control over trial conduct, protocol implementation, regulatory interactions, data collection, and analysis, reporting practices and compliance activities than we would in a traditional company-sponsored clinical trial. We may also have less firsthand knowledge regarding the conduct of the IIT and the generation, collection and analysis of data from the IIT. If data generated from the IIT are incomplete, inaccurate, unreliable or otherwise inadequate, or if we are unable to obtain such data on a timely basis, our ability to use the results of the IIT to inform the design and conduct of our planned Phase 1/2 clinical trial or otherwise advance the development of CITY-TFR2 could be adversely affected.
Clinical research conducted in China is subject to local laws, regulations, ethics committee approvals and governmental oversight, which may differ from those in the United States, Europe or other jurisdictions. The laws and regulations in China on therapies that are eligible for IITs in China are subject to differing interpretations by investigators, institutions, ethics committees, and regulators, and we may face delays in receiving, and may not receive, regulatory approval necessary to conduct the IITs in China, which may further delay and adversely affect our development of CITY-TFR2. Regulatory requirements, interpretations and enforcement practices may change, and we may experience delays in obtaining approvals, enrolling patients, collecting or transferring data, or completing the study. In addition, geopolitical tensions, trade restrictions, obligations related to data privacy and security, cross-border data transfer regulations or other governmental actions could adversely affect our
ability to oversee the trial, access trial data or use resulting data to support future regulatory submissions. Additionally, recent policy proposals in the United States may make acceptance by the FDA or inclusion in a marketing application of foreign data more difficult or costly.
If the IIT is delayed, suspended, terminated, conducted inconsistently with applicable requirements, generates incomplete, inaccurate, unreliable or otherwise inadequate data, produces data that Regulatory Authorities do not accept, raises safety concerns or results in AEs, or if the IIT does not generate clinical proof-of-concept data sufficient to support the continued development of CITY-TFR2 or inform our planned Phase 1/2 clinical trial, our development programs, regulatory strategy, business, financial condition, results of operations and prospects could be materially adversely affected.
Regulatory Authorities may not accept data from clinical trials conducted outside the United States, which could require us to conduct additional clinical trials and delay the development or approval of our product candidates.
As part of our global development strategy, some clinical trials are being conducted outside the United States, including our planned IIT for CITY-TFR2 in patients with anemia associated with myelofibrosis in China. We may conduct additional clinical trials for our current or future product candidates outside the United States. Although data from such trials may support our development programs, there can be no assurance that Regulatory Authorities will accept data generated outside their jurisdictions to support regulatory approval.
The acceptance of data from clinical trials conducted outside the United States or another jurisdiction by Regulatory Authorities may be subject to certain conditions or may not be accepted at all. In cases where data from foreign clinical trials are intended to serve as the basis for marketing approval in the United States, regardless of whether such trials were conducted under an IND, the FDA will generally not approve the application on the basis of foreign data alone unless the data are applicable to the United States population and United States medical practice, the trials were performed by clinical investigators of recognized competence and pursuant to GCP regulations, and the FDA can validate the data through on-site inspections or other appropriate means. The FDA has signaled that foreign GCP inspections will be increasing, alongside greater transparency of GCP inspection outcomes.
Many foreign Regulatory Authorities have similar approval requirements, including in relation to the use of data from clinical trials conducted in foreign jurisdictions. In addition, such foreign trials are subject to the applicable local laws of the foreign jurisdictions where the trials are conducted.
There can be no assurance that Regulatory Authorities will accept data from trials conducted outside the United States or the applicable jurisdiction. If Regulatory Authorities do not accept such data, including data from our planned IIT in China, we may be required to conduct additional clinical trials, which could be costly and time-consuming and could delay or prevent the development, regulatory approval or commercialization of our product candidates in the applicable jurisdiction. Additionally, recent policy proposals in the United States may make acceptance by the FDA or inclusion in a marketing application of foreign data more difficult or costly.
Where appropriate, we may seek approval from Regulatory Authorities through the use of expedited approval pathways, such as accelerated approval or comparable foreign abbreviated pathways. If we are unable to obtain such approvals, we may be required to conduct additional preclinical studies or clinical trials beyond those that we contemplate, which could increase the expense of obtaining, and delay the receipt of, necessary marketing approvals. Even if we receive accelerated approval from the FDA or approval following comparable foreign abbreviated pathways by foreign Regulatory Authorities, if our confirmatory trials do not verify clinical benefit, or if we do not comply with rigorous post-marketing requirements, the FDA or such Regulatory Authorities may seek to withdraw the accelerated approval.
Where appropriate, we may seek approval from Regulatory Authorities through the use of expedited approval pathways, such as accelerated approval or comparable foreign abbreviated pathways. There can be no assurance that we will decide to pursue, or that Regulatory Authorities will grant, such expedited approvals. Regulatory Authorities may require us to conduct additional preclinical studies or clinical
trials beyond those that we contemplate, which could increase the expense of obtaining, and delay the receipt of, necessary marketing approvals. Even if we receive accelerated approval from the FDA or approval following comparable foreign abbreviated pathways by foreign Regulatory Authorities, our approval may be subject to post-marketing requirements, including confirmatory trials, and may be withdrawn if such requirements are not satisfied or clinical benefit is not verified.
If granted, accelerated approval is usually contingent on the sponsor’s agreement to conduct, in a diligent manner, additional post-approval confirmatory studies to verify and describe the drug’s clinical benefit. Under the Food and Drug Omnibus Reform Act of 2022 (“FDORA”), the FDA is permitted to require, as appropriate, that a post-approval confirmatory study or studies be underway prior to approval or within a specified time period after the date of approval for a product granted accelerated approval. FDORA also gives the FDA increased authority to withdraw approval of a drug or biologic granted accelerated approval on an expedited basis if the sponsor fails to conduct such studies in a timely manner, send status updates on such studies to the FDA every 180 days to be publicly posted by the agency, or if such post-approval studies fail to verify the drug’s predicted clinical benefit. The FDA is empowered to act, such as issuing fines, against companies that fail to conduct with due diligence any post-approval confirmatory study or submit timely reports to the agency on their progress.
Prior to seeking accelerated approval, or approval following comparable foreign abbreviated pathways, we would seek feedback from Regulatory Authorities and would otherwise evaluate our ability to seek and receive such accelerated approval or approval following comparable foreign abbreviated pathways. There can be no assurance that after our evaluation of the feedback and other factors we will decide to pursue or submit an NDA for accelerated approval or any other form of expedited development, review or approval. Similarly, there can be no assurance that after subsequent feedback from Regulatory Authorities, we will continue to pursue or apply for accelerated approval or any other form of expedited development, review or approval, even if we initially decide to do so. Furthermore, if we decide to apply for accelerated approval, or comparable foreign abbreviated pathways, there can be no assurance that such application will be accepted or that any approval will be granted on a timely basis, or at all. Regulatory Authorities could also require us to conduct further studies prior to considering our application or granting approval of any type, including, for example, if other products are approved via the accelerated pathway, or comparable foreign abbreviated pathway, and subsequently converted by Regulatory Authorities to full approval. A failure to obtain accelerated approval or any other form of expedited development, review or approval for our product candidate would result in a longer period to commercialization of such product candidate, could increase the cost of development of such product candidate and could harm our competitive position in the marketplace.
Even if we obtain regulatory approval for a product candidate, we will be subject to ongoing regulatory obligations and continued regulatory review, which may result in significant additional expense and we may be subject to penalties if we fail to comply with regulatory requirements or experience unanticipated problems with our product candidates.
Even if our product candidates are approved, they will be subject to ongoing regulatory requirements for manufacturing, labeling, packaging, storage, advertising, promotion, sampling, record-keeping, conduct of post-marketing studies and submission of safety, efficacy and other post-market information, including both federal and state requirements in the United States and requirements of comparable foreign regulatory authorities. In addition, we will be subject to continued compliance with cGMP and GCP requirements for any clinical trials that we conduct post-approval. For example, the holder of an approved NDA is obligated to monitor and report AEs and any failure of a product to meet the specifications in the NDA. The holder of an approved NDA must also submit new or supplemental applications and obtain FDA approval for certain changes to the approved product, product labeling or manufacturing process. Advertising and promotional materials must comply with FDA rules and are subject to FDA review, in addition to other potentially applicable federal and state laws. If we fail to comply with applicable regulatory requirements following approval of any of our product candidates, a Regulatory Authority may: issue a FDA Form 483, or warning letter asserting that we are in violation of the law and potentially restricting our ability to sell, manufacture, import or export our products; seek an injunction or impose civil or criminal penalties or monetary fines; suspend or withdraw regulatory approval or revoke a license; suspend any ongoing clinical trials; refuse to approve a pending NDA or
supplements to an NDA submitted by us; seize product; or refuse to allow us to enter into supply contracts, including government contracts. Any government investigation of alleged violations of law could require us to expend significant time and resources in response and could generate negative publicity. The occurrence of any event or penalty described above may inhibit our ability to commercialize any approved products and generate revenues.
If we are successful in gaining approval for any of our product candidates, we and our CDMOs, which manufacture our products under contract, will continue to face significant regulatory oversight of the manufacturing and distribution of our products. Product manufacturers and their facilities are subject to payment of user fees and continual review and periodic inspections by Regulatory Authorities for compliance with cGMP and adherence to commitments made in the NDA. If we or a Regulatory Authority discovers previously unknown problems with a product such as AEs of unanticipated severity or frequency, or problems with the facility where the product is manufactured, a Regulatory Authority may impose restrictions relative to that product or the manufacturing facility, including requiring recall or withdrawal of the product from the market or suspension of manufacturing.
Any regulatory approvals that we receive for our product candidates may be subject to limitations on the approved indicated uses for which the product may be marketed or to the conditions of approval, or contain requirements for potentially costly post-marketing testing, including Phase 4 clinical trials and surveillance to monitor the safety and efficacy of the product candidate. Certain endpoint data we hope to include in any approved product labeling also may not make it into such labeling, including exploratory or secondary endpoint data such as patient-reported outcome measures. The FDA may also require a Risk Evaluation and Mitigation Strategy (“REMS”) as a condition of approval of our product candidates, which could entail requirements for long-term patient follow-up, a medication guide, physician communication plans or additional elements to ensure safe use, such as restricted distribution methods, patient registries and other risk minimization tools. Comparable requirements may apply in foreign countries. In addition, if Regulatory Authorities approves any of our product candidates, we will have to comply with requirements including submissions of safety and other post-marketing information, reports and registration.
Regulatory Authorities may impose consent decrees or withdraw or vary approval if compliance with regulatory requirements and standards is not maintained or if problems occur after the product reaches the market. Later discovery of previously unknown problems with our product candidates, including AEs of unanticipated severity or frequency, or with our CDMOs or manufacturing processes, or failure to comply with regulatory requirements, may result in revisions to the approved labeling to add new safety information, imposition of post-market studies or clinical trials to assess new safety risks or imposition of distribution restrictions or other restrictions under a REMS program or a comparable foreign program. Other potential consequences include, among other things:
•
restrictions on the marketing or manufacturing of our products, withdrawal of the product from the market or voluntary product recalls;
•
fines, FDA Form 483s, warning letters or holds on clinical trials;
•
refusal by Regulatory Authorities to approve pending applications or supplements to approved applications filed by us or suspension, variation or withdrawal of approvals;
•
product seizure, detention or refusal to permit the import or export of our product candidates;
•
total or partial suspension of production, distribution, manufacturing or clinical trials;
•
operating restrictions;
•
suspension of licenses; and
•
injunctions, fines or the imposition of civil or criminal penalties.
Additionally, Regulatory Authorities strictly regulate marketing, labeling, advertising and promotion of products that are placed on the market. Products may be promoted only for the approved indications and in accordance with the provisions of the approved label.
The policies of Regulatory Authorities may change and additional government regulations may be enacted that could prevent, limit or delay regulatory approval of our product candidates. In addition, the United States Supreme Court’s July 2024 decision to overturn established case law giving deference to Regulatory Authorities’ interpretations of ambiguous statutory language has introduced uncertainty regarding the extent to which the FDA’s regulations, policies and decisions may become subject to increasing legal challenges, delays and/or changes. We cannot predict the likelihood, nature or extent of government regulation that may arise from future legislation or administrative action, either in the United States or abroad. For more information, see the section titled “Business — Government Regulation.”
If we are slow or unable to adapt to changes in existing requirements or the adoption of new requirements or policies, or if we are not able to maintain regulatory compliance, we may lose any marketing approval that we may have obtained and we may not achieve or sustain profitability.
We are subject to significant regulatory oversight with respect to manufacturing our product candidates. The manufacturing facilities of our CDMOs or suppliers may not meet regulatory requirements. Failure to meet cGMP requirements set forth in regulations promulgated by Regulatory Authorities could result in significant delays in and costs of our products.
The manufacturing of therapeutics for clinical trials or commercial sale is subject to extensive regulation. Components of a finished product approved for commercial use or used in clinical trials must be manufactured in accordance with cGMP requirements. These regulations govern manufacturing processes and procedures, including recordkeeping, and the implementation and operation of quality systems to control and assure the quality of products and materials used in clinical trials. Poor control of the cGMP production processes can lead to product quality failures that can impact our ability to supply product, resulting in cost overruns and delays to clinical timelines, which could be extensive.
Such production process issues include, but are not limited to: critical deviations in the manufacturing process; facility and equipment failures; contamination of the product due to an ineffective quality control strategy; facility contamination as assessed by the facility and utility environmental monitoring program; ineffective process, equipment or analytical change management, resulting in failed lot release criteria; raw material failures due to ineffective supplier qualification or regulatory compliance issues at critical suppliers; ineffective product stability; failed lot release or facility and utility QC testing; ineffective corrective actions or preventative actions taken to correct or avoid critical deviations due to our developing understanding of the manufacturing process as we scale; and failed or defective components or consumables.
We must supply all necessary documentation in support of an NDA or other marketing authorization application on a timely basis and must adhere to the cGMP requirements of Regulatory Authorities which are enforced, in the case of the FDA, in part through its facilities inspection program.
Regulatory authorities typically require representative manufacturing site inspections to assess adequate compliance with cGMPs and manufacturing controls as described in the filing. If either we or one of our third-party manufacturing sites fails to provide sufficient quality assurance or control, the product approval to commercialize may not be granted. Inspections by regulatory authorities may occur at any time during the development or commercialization phase of products. The inspections may be product specific or facility specific for broader cGMP inspections or as a follow up to market or development issues that the Regulatory Authority may identify. Deficient inspection outcomes may influence the ability of our CDMOs or suppliers to fulfill their supply obligations, impacting or delaying supply or delaying product candidates.
The manufacturing process for any products that we may develop is subject to the approval process of Regulatory Authorities, and we will need to contract with CDMOs which we believe can meet such requirements on an ongoing basis. If we or our CDMOs are unable to reliably produce product candidates to specifications acceptable to Regulatory Authorities, we or our collaboration partners may not obtain or maintain the approvals we or they need to commercialize such products. Even if we or our collaboration partners obtain regulatory approval for any of our product candidates, there is no assurance that either we or our CDMOs will be able to manufacture the approved product to
specifications acceptable to Regulatory Authorities, to produce it in sufficient quantities to meet the requirements for the potential launch of the product, or to meet potential future demand. Any of these challenges could delay completion of clinical trials, require bridging clinical trials or the repetition of one or more clinical trials, increase clinical trial costs, delay approval of our product candidates, impair commercialization efforts or increase our cost of goods. The occurrence of any of the foregoing could have an adverse effect on our business, financial condition, results of operations and prospects.
We have limited control over the manufacturing process of, and are dependent on, our CDMOs for compliance with cGMPs. If our CDMOs cannot successfully manufacture material that conforms to our specifications and the strict regulatory requirements of Regulatory Authorities, we may not be able to secure and/or maintain regulatory approval for our product candidates manufactured at these facilities. In addition, we have limited control over the ability of our CDMOs to maintain adequate quality control, quality assurance and qualified personnel. Furthermore, all of our CDMOs are engaged with other companies to supply or manufacture materials or products for such companies, which exposes our CDMOs to regulatory risks for the production of such materials and products. As a result, failure to meet the regulatory requirements for the production of those materials and products may generally affect the regulatory status of our CDMOs’ facility. Our failure, or the failure of our CDMOs, to comply with applicable regulations could result in sanctions being imposed on us, including clinical holds, fines, injunctions, civil penalties, delays, suspension or withdrawal of approvals, license revocation, seizures or recalls of product candidates or products, operating restrictions and criminal prosecutions, any of which could significantly and adversely affect supplies of our products and product candidates (including those of our collaboration partners) and our overall business operations. Our dependence upon others for the manufacture of our product candidates and raw materials may adversely affect our future profit margins and our ability to commercialize any products that receive regulatory approval on a timely and competitive basis.
We also may encounter problems hiring and retaining the experienced scientific, quality-control and manufacturing personnel needed to operate our manufacturing processes and operations, which could result in delays in production or difficulties in maintaining compliance with applicable regulatory requirements. While we will train and qualify all personnel around the appropriate handling of our products and materials, we may not be able to control or ultimately detect intentional sabotage or negligence by any employee or contractor.
Disruptions at the FDA, the SEC and other United States government agencies caused by reduction in staffing, leadership changes, funding shortages or global health concerns could hinder their ability to hire, retain or deploy key leadership and other personnel, or otherwise prevent new or modified products from being developed, approved or commercialized in a timely manner or at all, or otherwise prevent those agencies from performing normal business functions on which the operation of business may rely, which could negatively impact our business.
Federal agencies in the United States, including the FDA, the SEC and other comparable regulatory authorities, operate pursuant to annual appropriations and other political and budgetary processes, and may from time to time be subject to continuing resolutions, funding lapses, or other fiscal constraints. Currently, although the FDA and many other federal agencies have funding sufficient to continue their activities through December 11, 2026, the timing and amount of future funding is unpredictable. Without appropriation of sufficient funding to federal agencies, our business operations related to our product development activities for the United States market could be impacted. The ability of Regulatory Authorities to review and approve new products can be affected by a variety of factors, including government budget and funding levels, ability to hire and retain key personnel and accept the payment of user fees, statutory, regulatory, and policy changes, and other events that may otherwise affect Regulatory Authorities’ ability to perform routine functions. Average review times at Regulatory Authorities have fluctuated in recent years and may continue to fluctuate as a result of these factors. In addition, government funding of the SEC and other United States government agencies on which our operations may rely, including those that fund research and development activities, is subject to the political process, including executive and congressional priorities, which is inherently fluid and unpredictable. For example, the current administration has issued executive orders seeking to greatly reduce the size
of the federal workforce, including through layoffs and severance packages offered to employees of federal agencies within the executive branch and independent agencies, including the FDA. Any such reduction in personnel may result in longer review times by the FDA and other agencies.
Disruptions at the FDA, the SEC and other agencies, including as a result of substantial leadership changes, personnel cuts, policy changes or otherwise, may also slow the time necessary for new drugs to be reviewed and/or approved by necessary government agencies, which would adversely affect our business. Changes and cuts in FDA staffing have been reported as resulting in delays in the FDA’s responsiveness or in its ability to review IND submissions or marketing applications, issue regulations or guidance, or implement or enforce regulatory requirements in a timely fashion or at all. Also, state governments may seek to address or react to changes at the federal level with changes to their regulatory frameworks in a manner that could impact our operations.
Over the last several years, including from October 1, 2025 to November 12, 2025, the United States government shut down several times and certain regulatory agencies, such as the FDA and the SEC, have had to furlough critical employees and cease critical activities.
If a prolonged government shutdown occurs again, or a widespread freeze on federal funding occurs in the future or if global health concerns or staffing changes prevent the FDA, the SEC, or other comparable foreign regulatory authorities from conducting their regular inspections, reviews, or other regulatory activities, including formal and informal interactions with product developers, it could significantly impact the ability of the FDA, the SEC, or other such comparable foreign regulatory authorities to timely review and process our regulatory submissions, which could have a material adverse effect on our business. Further, in our operations as a public company, future government shutdowns and/or substantial leadership, personnel, and policy changes at the SEC could impact our business by delaying review of our public filings, which in turn could delay or frustrate our ability to access the public markets and obtain necessary capital in order to properly capitalize and continue our operations or delay the review or effectiveness of required regulatory or securities filings.
Healthcare legislative reform discourse and potential or enacted measures may have a material adverse impact on our business and results of operations and legislative or political discussions surrounding the desire for and implementation of pricing reforms may adversely impact our business.
In the United States and some foreign jurisdictions, there have been a number of legislative and regulatory changes and proposed changes regarding the healthcare system that could prevent or delay marketing approval of our product candidates, restrict or regulate post-approval activities and affect our ability to profitably sell any product candidates for which we obtain marketing approval. The pharmaceutical industry has been a particular focus of these efforts and has been significantly affected by legislative initiatives. Current laws, as well as other healthcare reform measures that may be adopted in the future, may result in more rigorous coverage criteria and in additional downward pressure on the price that we receive for any FDA-approved product. If reimbursement of our products is unavailable or limited in scope, our business could be materially harmed. For more information, please see the sections titled “Business — Government Regulation — Coverage and Reimbursement” and “— Healthcare Reform and Other Regulatory Changes” included elsewhere in this prospectus.
Third-party payors, whether domestic or foreign, or governmental or commercial, are developing increasingly sophisticated methods of controlling healthcare costs. In the United States and certain other jurisdictions, there have been, and are expected to continue to be, a number of legislative and regulatory changes to the healthcare system that could impact our ability to sell our products profitably. In addition, significant uncertainty exists in the United States and certain other jurisdictions regarding the provision and financing of healthcare because the elected administrations in such countries have publicly declared their intention to review and potentially significantly modify the current legal and regulatory framework for the healthcare system including through drug pricing and reimbursement reforms.
We cannot predict the initiatives that may be adopted in the future. The continuing efforts of the government, insurance companies, managed care organizations and other payers of healthcare services to contain or reduce costs of healthcare may adversely affect:
•
the demand for any of our product candidates, if approved;
•
the ability to set a price that we believe is fair for any of our product candidates, if approved;
•
our ability to generate revenues and achieve or maintain profitability;
•
the level of taxes that we are required to pay; and
•
the availability of capital.
For example, recent CMS drug pricing reforms, including the finalized Global Benchmark for Efficient Drug Pricing Model (“GLOBE”) for Medicare Part B, the proposed Guarding United States Medicare Against Rising Drug Costs (“GUARD”) model for Medicare Part D, and the GENErating cost Reductions fOr United States Medicaid (“GENEROUS”) Model, if implemented as currently proposed or finalized, could materially impact our revenue. Any reduction in reimbursement from Medicare or other government programs may result in a similar reduction in payments from private payors, which may adversely affect our future profitability and could materially reduce net realized prices.
Further, the Inflation Reduction Act of 2022 (the “IRA”) was recently revised to broaden the exemption from the drug price negotiation program for drugs with orphan designations. Previously under the IRA, orphan drugs were exempted from the Medicare drug price negotiation program; but this exemption was restricted to drugs with only one orphan designation and for which the only approved indication is for that rare disease or condition. If a product were to receive multiple orphan designations or had multiple approved indications, it would not have qualified for the orphan drug exemption. Under the One Big Beautiful Bill Act of 2025 (the “OBBBA”), this restriction was eliminated; and effective for the 2028 initial price applicability year, all orphan drugs, regardless of the number of orphan designations or indications, are exempt from the Medicare drug price negotiation program, provided that all approved indications are for rare diseases.
In December 2021, Regulation No 2021/2282 on Health Technology Assessment (“HTA”) amending Directive 2011/24/EU (the “Regulation”), was adopted in the EU. This Regulation, which entered into force in January 2022 and became applicable in January 2025, is intended to boost cooperation among Member States in assessing health technologies, including new medicinal products, and provides the basis for cooperation at the EU level for joint clinical assessments in these areas. The Regulation permits Member States to use common HTA tools, methodologies and procedures across the EU, working together in four main areas, including joint clinical assessment of the innovative health technologies with the most potential impact for patients, joint scientific consultations whereby developers can seek advice from HTA authorities, identification of emerging health technologies to identify promising technologies early, and continuing voluntary cooperation in other areas. Individual Member States will continue to be responsible for assessing nonclinical (e.g., economic, social, ethical) aspects of health technologies and making decisions on pricing and reimbursement.
Any of the foregoing measures (and related implementing guidance, rulemaking, enforcement activity or private litigation) could materially reduce pricing flexibility, increase rebates or discounts, narrow coverage, delay patient access, and adversely affect our ability to commercialize any current or future product candidates, if approved.
Risks Related to Our Reliance on Third Parties
We rely on a limited number of third parties for the supply and manufacture of our product candidates for our research, preclinical and clinical activities, and may do the same for commercial supplies of our products, if approved. As our pipeline increases and matures, the increased demand for supplies from our manufacturers may increase the risk that we will not have sufficient supply when needed or at an acceptable cost.
We currently utilize, and expect to continue to utilize, a limited number of CDMOs to, among other things, supply and manufacture raw materials, intermediates, drug substance and drug product, perform analytical and quality testing and supply materials for our preclinical and clinical supply for all of our product candidates. For example, we are party to manufacturing and supply agreements with certain
CDMOs to support the manufacture and supply of our product candidates. Given the specialized expertise, manufacturing capabilities and capacity required to manufacture RNAi therapeutics, we expect to continue to rely on third-party CDMOs for significant aspects of the manufacture and supply of our product candidates. Although we may qualify additional manufacturers over time, there can be no assurance that our existing or future CDMOs will be able to meet our manufacturing requirements on acceptable terms or at all.
In order to produce sufficient quantities to meet the demand for clinical trials and, if approved, subsequent commercialization of our product candidates, our CDMOs will be required to increase their production and optimize their manufacturing processes while maintaining the quality, consistency and regulatory compliance of our product candidates, as applicable. The transition to larger scale production could prove difficult. If our third-party manufacturers are not able to optimize their manufacturing processes to increase the product yield for our product candidates, or if they are unable to produce increased amounts of our product candidates while maintaining the same quality, then we may not be able to meet the demands of clinical trials or market demands, which could adversely impact our ability to timely conduct our clinical trials or commercialize our product candidates, if approved, and could have a material adverse impact on our business and results of operations. Furthermore, as the number of companies developing RNAi therapeutics and other oligonucleotide-based medicines continues to increase, competition for specialized manufacturing capacity, raw materials, equipment and other resources necessary to manufacture our product candidates may also increase. Any shortage of, or delay in obtaining, such manufacturing capacity or materials could increase our manufacturing costs, delay the manufacture or supply of our product candidates, and adversely affect our ability to advance our development programs or commercialize our product candidates, if approved.
Even if we are able to maintain arrangements with our CDMOs, reliance on CDMOs entails additional risks, including:
•
the failure of the CDMO to comply with applicable regulatory requirements and reliance on third parties for manufacturing process development, regulatory compliance and quality assurance;
•
manufacturing delays if our CDMOs give greater priority to the supply of other products over our product candidates or otherwise do not perform satisfactorily according to the terms of the agreement between us;
•
limitations on supply availability resulting from capacity and scheduling constraints of third parties;
•
the possible breach of manufacturing agreements by our CDMOs because of factors beyond our control;
•
the possible termination or non-renewal of the manufacturing agreements by our CDMOs, at a time that is costly or inconvenient to us; and
•
the possible misappropriation of our proprietary technology and intellectual property, including our know-how.
If we are unable to maintain our key manufacturing relationships, we may fail to find replacement CDMOs, which could delay or impair our ability to obtain regulatory approval for our product candidates. If we do find replacement CDMOs, we may not be able to enter into agreements with them on terms and conditions favorable to us and there could be a substantial delay before new facilities could be qualified and registered with Regulatory Authorities.
We do not currently have long-term supply contracts with all of our suppliers and they are not obligated to supply materials to us for any period, in any specified quantity or at any certain price beyond the delivery contemplated by the relevant purchase orders. As a result, our suppliers could stop selling to us at commercially reasonable prices, or at all. While we intend to enter into long-term master supply agreements with certain of our suppliers and manufacturers in the future as we advance our clinical trials or commercialization plans, we may not be successful in negotiating such agreements on favorable terms or at all. Our failure to secure these arrangements as needed could have a material
adverse effect on our ability to complete the development of our product candidates or, to commercialize them, if approved. If we do enter into such long-term master supply agreements, or enter into such agreements on less favorable terms than we currently have with such manufacturers, we could be subject to binding long-term purchase obligations that may be harmful to our business, including in the event that we do not conduct our trials on planned timelines or utilize the materials that we are required to purchase.
Additionally, if CDMO with whom we contract fails to perform its obligations, we may be forced to manufacture the materials ourselves, for which we may not have the capabilities or resources, or enter into an agreement with a different CDMO. In either scenario, our clinical trials supply could be delayed significantly as we establish alternative supply sources. In some cases, the technical skills required to manufacture our product candidates may be unique or proprietary to the original CDMO and we may have difficulty, or there may be contractual restrictions prohibiting us from transferring such skills to a back-up or alternate supplier, or we may be unable to transfer such skills at all. In addition, if we are required to change CDMOs for any reason, we will be required to verify that the new CDMO maintains facilities and procedures that comply with quality standards and with all applicable regulations. We will also need to verify, such as through a manufacturing comparability study, that any new manufacturing process will produce our product candidates according to the specifications previously submitted to Regulatory Authorities. We may be unsuccessful in demonstrating the comparability of clinical supplies, which could require the conduct of additional clinical trials. The delays associated with the verification of a new CDMO could negatively affect our ability to develop or commercialize our product candidates in a timely manner or within budget. Furthermore, a CDMO may possess technology related to the manufacture of our product candidates that such third party owns independently. This would increase our reliance on such CDMO or require us to obtain a license from such CDMO in order to have another third party manufacture our product candidates.
If any of our product candidates are approved by any Regulatory Authority, we will likely utilize arrangements with CDMOs for the commercial production of such product. This process is difficult and time-consuming and we may face competition for access to manufacturing facilities as there are a limited number of CDMOs operating under cGMPs that are capable of manufacturing our product candidates. Consequently, we may not be able to reach agreement with CDMOs on satisfactory terms, which could delay our commercialization.
The operations of our suppliers and CDMOs, some of which are located outside of the United States, are subject to additional risks that are beyond our control and that could harm our business, financial condition, results of operations and prospects. As a result of our global suppliers, we are subject to risks associated with doing business abroad, including:
•
political unrest, terrorism, labor disputes and economic instability resulting in the disruption of trade from foreign countries in which our products are manufactured;
•
the imposition of new laws and regulations, including those relating to labor conditions, quality, and safety standards, imports, duties, taxes and other charges on imports, as well as trade restrictions and restrictions on currency exchange or the transfer of funds, particularly new or increased tariffs imposed on imports from countries where our suppliers operate;
•
greater challenges and increased costs with enforcing and periodically auditing or reviewing our suppliers’ and CDMOs’ compliance with cGMPs or status acceptable to Regulatory Authorities;
•
reduced protection for intellectual property rights, including trademark protection, in some countries;
•
disruptions in operations due to global, regional or local public health crises or other emergencies or natural disasters;
•
disruptions or delays in shipments; and
•
changes in local economic conditions in countries where our CDMOs or suppliers are located.
In particular, there is currently significant uncertainty about the future relationship between the United States and various other countries, including countries in which our suppliers and CDMOs operate, with respect to trade policies, treaties, government regulations and tariffs. It is possible that further tariffs or other trade restrictions may be imposed that could affect imports of APIs or other materials used in our product candidates, or otherwise disrupt our supply chain or increase our manufacturing costs. Given the evolving global trade environment and uncertainty regarding how the United States or foreign governments will act with respect to tariffs, international trade agreements and policies, further governmental action related to tariffs, additional taxes, contracting matters, regulatory changes or other retaliatory trade measures could occur and have a material adverse effect on our business, financial condition, results of operations and prospects. These and other factors beyond our control could interrupt our suppliers’ and CDMOs’ production, influence their ability to manufacture or supply our clinical materials cost-effectively or at all, or inhibit their ability to procure necessary raw materials, any of which could adversely affect our business, financial condition, results of operations and prospects.
We depend on limited source suppliers for certain drug substances, drug products, raw materials, samples, components and other materials used in our product candidates. If we are unable to source these supplies on a timely basis, or establish longer-term contracts with our suppliers, we may not be able to complete our clinical trials on time and the development of our product candidates may be delayed.
We depend on limited source suppliers for certain raw materials, APIs, drug products, drug substances and other materials used in our product candidates. For example, we rely on a limited number of CDMOs, including in China, to provide certain manufacturing and supply services for our product candidates. Although we may qualify additional suppliers over time, certain materials, manufacturing processes and services are currently available from only a limited number of qualified suppliers. Any disruption in the supply of these materials or services, any inability of our suppliers to meet our manufacturing requirements, or any change in our relationships with these suppliers or the contractual terms governing such relationships could adversely affect our business, financial condition, results of operations and prospects. Moreover, there may be difficulties in scaling the manufacture of our product candidates for later-stage clinical development or commercialization, and the costs of manufacturing could become prohibitive.
Furthermore, any of the limited source suppliers upon whom we rely could stop producing our supplies, cease operations or be acquired by, or enter into exclusive arrangements with, our competitors. In addition, geopolitical tensions may impact our suppliers, some of which are located outside of the United States, including in China. There is currently significant uncertainty about the future relationship between the United States and various other countries, including China, with respect to trade policies, treaties, government regulations and tariffs. Increased tariffs or pending legislation that would impose federal contracting or federal funding limitations on parties directly using or connected to those using the services or equipment of certain foreign entities with known or alleged associations with foreign adversaries could potentially disrupt our existing supply chains and impose additional costs on our business. For example, the BIOSECURE Act, enacted on December 18, 2025 as part of the National Defense Authorization Act for Fiscal Year 2026, restricts United States federal agencies from entering into contracts, grants, or loans with entities that use biotechnology equipment or services provided by designated “biotechnology companies of concern” (“BCCs”). BCCs are identified through designation by the Office of Management and Budget (“OMB”) following a multi-agency review process, or through inclusion on the Department of Defense’s annual list of Chinese military companies maintained pursuant to Section 1260H of the National Defense Authorization Act (“NDAA”) for Fiscal Year 2021 (the “1260H List”), subject to an OMB determination that the listed entity is involved in the manufacture, distribution, provision, or procurement of biotechnology equipment or services. Certain CDMOs we engage are not currently designated as BCCs under either pathway; however, there is no assurance that any such CDMO will not be added to the 1260H List or otherwise designated as a BCC in future OMB determinations. The Act’s prohibitions are not yet in effect, and OMB must first publish a formal BCC list (due no later than December 2026), after which OMB has 180 days to issue guidance regarding the Act, and then the Federal Acquisition Regulatory Council has one year to revise the Federal Acquisition Regulations, with prohibitions taking effect 60 or 90 days thereafter (depending on how the entity is
designated as a BCC). Under the statute, restrictions for newly designated BCCs may not take effect until 2027 or later. The Act also provides a five-year grandfathering period for biotechnology equipment or services provided under contracts with a BCC entered into prior to the applicable effective date. To the extent we receive or in the future receive federal funding — including grants, loans, or contracts subject to the Federal Acquisition Regulations — in connection with which any CDMO we engage is or becomes designated a BCC, we could be required to transition to alternative manufacturers or risk ineligibility for such funding. Even absent a direct federal nexus, the designation of any CDMO we engage as a BCC, or the prospect of such designation, could disrupt our supply chain, impair such CDMO’s ability to operate or attract United States customers, or require us to identify and qualify an alternative sole-source manufacturer, any of which could cause significant delays in our clinical development program and have a material adverse effect on our business, financial condition, and results of operations. If any third party that we engage to supply any materials or manufacture products for our preclinical studies and clinical trials should cease to continue to do so, or if we are prevented from using their services for any reason, we could experience delays in advancing these studies and trials while we identify and qualify replacement suppliers.
Establishing additional or replacement suppliers, and obtaining regulatory clearance or approvals that may result from adding or replacing suppliers, could take a substantial amount of time, result in increased costs and impair our ability to produce our products, which would adversely impact our business, financial condition, results of operations and prospects. Any such interruption or delay may force us to seek similar supplies from alternative sources, which may not be available at reasonable prices, or at all. Any interruption in the supply of sole source or limited source components for our product candidates would adversely affect our ability to meet scheduled timelines and budget for the development and commercialization of our product candidates, could result in higher expenses and would harm our business. Although we have not experienced any significant disruption as a result of our reliance on limited or sole source suppliers, we have a limited operating history and cannot assure you that we will not experience disruptions in our supply chain in the future as a result of such reliance or otherwise.
We rely on and expect to continue to rely on third parties to conduct aspects of our research, preclinical studies, clinical protocol development and clinical trials for our programs and product candidates. If these third parties do not perform satisfactorily, comply with regulatory requirements or meet expected deadlines, we may not be able to develop product candidates in a timely or cost-effective manner, or obtain regulatory approval for or commercialize our product candidates and our business could be substantially harmed.
We currently rely and expect to continue to rely on third parties, such as CROs, medical institutions and clinical investigators, to conduct our clinical trials, including our planned IIT for CITY-TFR2 in anemia in China. We currently rely and expect to continue to rely on third parties to conduct certain research and preclinical testing activities. In some cases, these third parties may terminate their engagements with us. If we need to enter into alternative arrangements, it could delay our product development activities or increase our costs.
Our reliance on these third parties for research and development activities will reduce our control over these activities but will not relieve us of our regulatory or contractual responsibilities. We will be responsible for ensuring that each of our preclinical studies and clinical trials is conducted in accordance with the applicable protocol, legal and regulatory requirements and scientific standards. For example, we will remain responsible for ensuring that each of our clinical trials is conducted in accordance with the general investigational plan and protocols for the trial. Moreover, the FDA requires us to comply with regulations, commonly referred to as GCPs for conducting, recording and reporting the results of clinical trials to assure that data and reported results are credible and accurate and that the rights, integrity and confidentiality of trial participants are protected. We also are required to register ongoing clinical trials and post the results of completed clinical trials on a government-sponsored database, ClinicalTrials.gov, within certain timeframes for publication. Failure to do so can result in fines, adverse publicity and civil and criminal sanctions. For any violations of laws and regulations during the conduct of our preclinical studies and clinical trials, we could be subject to warning letters or enforcement action that may include civil penalties up to and including criminal prosecution.
We and our CROs will be required to comply with regulations, including GCPs, for conducting, monitoring, recording and reporting the results of preclinical studies and clinical trials to ensure that the data and results are scientifically credible and accurate and that the trial participants are adequately informed, among other things, of the potential risks of participating in clinical trials. We are also responsible for ensuring that the rights of our clinical trial participants are protected. These regulations are enforced by Regulatory Authorities for any product candidates in clinical development. The FDA enforces GCP regulations through periodic inspections of clinical trial sponsors, principal investigators and trial sites. If we or our CROs fail to comply with applicable GCPs, the clinical data generated in our clinical trials may be deemed unreliable and Regulatory Authorities may require us to perform additional clinical trials before approving our marketing applications. There is no assurance that the FDA or other Regulatory Authorities, upon inspection, will determine that any of our future clinical trials will comply with GCPs. In addition, our clinical trials must be conducted with product candidates produced in accordance with the requirements in cGMP regulations. Our failure or the failure of our CROs to comply with these regulations may require us to repeat clinical trials, which would delay the regulatory approval process and could also subject us to enforcement action.
Although we intend to design the clinical trials for certain of our product candidates, our collaboration partners may design the clinical trials that they are managing (in some cases, with our input) and in the case of clinical trials controlled by us, we expect that CROs will perform many of the activities required to conduct clinical trials. As a result, many important aspects of our development programs, including their conduct and timing, will, in many respects, be outside of our direct control. Our reliance on third parties to conduct or manage our current or future preclinical studies and clinical trials will also result in less direct control over the management of data developed through preclinical studies and clinical trials than would be the case if we were relying entirely upon our own staff. Communicating with outside parties can also potentially lead to mistakes as well as difficulties in coordinating activities. Outside parties may: have staffing difficulties; fail to comply with contractual obligations; experience regulatory compliance issues; undergo changes in priorities or become financially distressed; form relationships with other entities, some of which may be our competitors; have human errors or be subject to cyber-attacks.
These factors may materially adversely affect the willingness or ability of third parties to conduct our preclinical studies and clinical trials and may subject us to unexpected cost increases that are beyond our control. If the CROs do not perform preclinical studies and clinical trials in a satisfactory manner, breach their obligations to us or fail to comply with regulatory requirements, the development, regulatory approval and commercialization of our product candidates may be delayed, we may not be able to obtain regulatory approval and commercialize our product candidates, or our development programs may be materially and irreversibly harmed. If we are unable to rely on preclinical and clinical data collected by our CROs, we could be required to repeat, extend the duration of or increase the size of any clinical trials we conduct and this could significantly delay commercialization and require significantly greater expenditures.
We also expect to rely on other third parties to transport, store and distribute the required materials for our clinical trials. Any performance failure on the part of these third parties could result in damaged products and could delay clinical development or marketing approval of any product candidates we may develop or commercialization of our products, if approved, producing additional losses and depriving us of potential product revenue, causing us to default on our contractual commitments, result in losses that are not covered by insurance, and damage our reputation and overall perception of our products in the marketplace.
Any of the third party organizations we utilize may terminate their engagements with us under certain circumstances. The replacement of an existing CRO or other third party may result in the delay of the affected trials or otherwise adversely affect our efforts to obtain regulatory approvals and commercialize our product candidates. For example, although we believe there are a number of other CROs we could engage, we may not be able to enter into alternative arrangements or do so on commercially reasonable terms. In addition, while we believe there may be suitable replacements for one or more of these service providers, there is a natural transition period when a new service provider
begins work. As a result, delays may occur, which could negatively impact our ability to meet our expected clinical development timelines and harm our business, financial condition, results of operations and prospects.
We have in the past entered into, and in the future may enter into, partnership, collaboration and licensing arrangements with third parties to support development of programs and product candidates. If these partnership, collaboration and licensing arrangements are not successful, our business could be adversely affected.
We have entered into or sought to enter into partnership, collaboration and licensing arrangements with third parties, which we refer to generally as our “collaboration partners” for strategic purposes, including for purposes of collaborating with collaboration partners with distinctive capabilities or experience with different modalities, working with collaboration partners capable of advancing the development and commercialization of our product candidates, and providing access to additional capital.
For example, in addition to our wholly owned pipeline, we have entered into research collaboration and license agreements with two industry collaboration partners. In July 2024, we entered into the B+L Agreement with Bausch + Lomb focused on the discovery and development of RNAi therapeutics for ocular diseases, and in May 2025, we entered into the Biogen Agreement with Biogen focused on the discovery and development of an RNAi program for neurodegenerative diseases. In addition, in October 2023, we entered into the OSIF License Agreement with OSIF, pursuant to which OSIF granted us an exclusive, royalty-bearing license to certain patent rights relating to the originating technology underlying cityRNAs and certain other aspects of our technology platform. We expect to enter into additional partnership, collaboration and licensing arrangements in the future. Our existing arrangements, as well as any future arrangements we may enter into, could pose a number of risks, including the following:
•
collaboration partners may not perform their obligations as expected;
•
collaboration partners may not commit sufficient resources, including personnel, capital, and management attention, to the development and potential commercialization of the applicable product candidates;
•
collaboration partners may change their strategic focus, undergo a change of control, or otherwise deprioritize our collaboration programs in favor of other programs or therapeutic areas;
•
the clinical trials conducted as part of such partnership, collaboration and licensing arrangement may not be successful;
•
collaboration partners may not pursue development and commercialization of any product candidates that achieve regulatory approval or may elect not to continue or renew development or commercialization of programs based on clinical trial results, changes in the strategic collaborators’ focus or available funding, or external factors, such as an acquisition, which divert resources or create competing priorities;
•
collaboration partners may delay clinical trials, provide insufficient funding for clinical trials, stop a clinical trial, abandon a product candidate, repeat or conduct new clinical trials or require a new formulation of a product candidate for clinical testing;
•
collaboration partners could independently develop, or develop with third parties, products that compete directly or indirectly with our product candidates if the strategic collaborators believe that competitive products are more likely to be successfully developed or can be commercialized under terms that are more economically attractive than ours;
•
product candidates developed in partnership, collaboration and licensing arrangements with us may be viewed by our collaboration partners as competitive with their own candidates or products, which may cause collaboration partners to cease to devote resources to the development of our programs or the development or commercialization of our product candidates;
•
a collaboration partner with marketing and distribution rights to one or more of our product candidates that achieve regulatory approval may not commit sufficient resources to the marketing and distribution of any such product;
•
disagreements with collaboration partners, including disagreements over proprietary rights, contract interpretation or the preferred course of development of any product candidates, may cause delays or termination of the research, development or commercialization of such product candidates, may lead to additional responsibilities for us with respect to such product candidates or may result in litigation or arbitration, any of which would be time-consuming and expensive;
•
collaboration partners may not properly maintain or defend our intellectual property rights or may use our proprietary information in such a way as to invite litigation that could jeopardize or invalidate our intellectual property or proprietary information or expose us to potential litigation;
•
disputes may arise with respect to the interpretation of key terms regarding control, economic rights, or the ownership of intellectual property developed pursuant to our partnership, collaboration and licensing arrangements;
•
collaboration partners may infringe the intellectual property rights of third parties, which may expose us to litigation and potential liability;
•
partnership, collaboration and licensing arrangements may, in certain instances, be terminated for the convenience of the collaboration partner and, if terminated, the development of our programs and product candidates may be delayed, or we may lose rights to intellectual property or expertise related to such programs and product candidates, and we could be required to raise additional capital to pursue further development or commercialization of the applicable product candidates;
•
future relationships may require us to incur non-recurring and other charges, assume indebtedness or contingent liabilities, increase our near- and long-term expenditures, acquire intangible assets, issue securities that dilute our existing stockholders, disrupt our management and business, or otherwise impact our ability to generate revenue from acquired intellectual property, technology and/or products sufficient to meet our objectives or even to offset the associated transaction and maintenance costs;
•
we could face significant competition in seeking appropriate collaboration partners and the negotiation process and diligence process is time-consuming and complex; and
•
our international operations, through any future partnerships, collaborations, acquisitions or joint ventures, may expose us to certain operating, legal, and other risks not encountered in the United States.
Whether we reach a definitive agreement for a partnership, collaboration or licensing arrangement will depend, among other things, on our assessment of the collaboration partner’s resources and expertise, the terms and conditions of the proposed partnership, collaboration or licensing arrangement, and the potential collaboration partner’s evaluation of a number of factors. Those factors may include, among others: (i) our technologies and capabilities; (ii) our intellectual property position with respect to the subject program or product candidate; (iii) the design or results of clinical trials; (iv) the likelihood of approval by Regulatory Authorities; (v) the potential market for the subject product candidate; (vi) potential competing products; and (vii) industry and market conditions generally. In addition, the significant number of business combinations among large pharmaceutical and biotechnology companies has reduced the number of potential future collaboration partners with whom we can partner.
Partnerships, collaborations and licensing arrangements are complex and time-consuming to negotiate and document. We may have to relinquish valuable rights to our programs and product candidates, intellectual property or future revenue streams, or grant licenses on terms that are not favorable to us or in instances where it would have been more advantageous for us to retain sole development and commercialization rights. For some programs and product candidates, we depend on collaboration partners to design and conduct the clinical trials. As a result, we may not control the
manner or time schedule in which these clinical trials are conducted, which may negatively impact our business operations. In addition, if any of our collaboration partners withdraws support for one or more of our programs or product candidates or otherwise impairs their development, our business could be negatively affected. In addition, management of our relationships with collaboration partners requires (i) significant time and effort from our management team; (ii) coordination of our marketing and research and development programs with the marketing and research and development priorities of our collaborators and (iii) effective allocation of our resources across multiple projects.
Partnerships, collaborations and licensing arrangements may never result in the successful development of programs or development and commercialization of product candidates or the generation of sales revenue. The success of these arrangements will depend heavily on the efforts and activities of our collaboration partners. Collaboration partners generally have significant discretion in determining the efforts and resources that they will apply to the development of programs and the development and commercialization of product candidates, and they may not pursue or prioritize the development and commercialization of such programs and product candidates in a manner that is in our best interests. Product revenues arising from partnership, collaboration and licensing arrangements are likely to be lower than if we directly marketed and sold products. Disagreements with collaboration partners regarding clinical development or commercialization matters can lead to delays in the development process or commercialization of the applicable product candidate and, in some cases, the termination of the partnership, collaboration or licensing arrangement. These disagreements can be difficult to resolve if neither of the parties has final decision-making authority. Partnership, collaboration and licensing arrangements are often terminable by the collaboration partner, and any such termination or expiration would adversely affect us financially and could harm our business reputation. If we were to become involved in arbitration or litigation with any of our collaboration partners, it would consume time and divert management resources away from operations, damage our reputation, impact our ability to enter into future partnership, collaboration and licensing arrangements and may further result in substantial payments from us to our collaboration partners to settle those disputes.
We may not be able to establish additional partnership, collaboration and licensing arrangements on a timely basis, on acceptable terms, or at all, or to maintain and successfully conclude such arrangements. Such arrangements with third parties could cause us to expend significant resources and incur substantial business risk with no assurance of financial return. In addition, given the nature of our relationships with our collaboration partners, we often do not fully control the progression, clinical development, regulatory strategy or eventual commercialization, if approved, of our jointly developed product candidates. As a result, our future success and the potential to receive revenues under these partnership, collaboration and licensing arrangements are significantly dependent on our collaboration partners’ efforts, over which we have little control. If we are unable to establish or maintain partnership, collaboration and licensing arrangements on terms favorable to us, if our partnership, collaboration and licensing arrangements do not result in the successful development and commercialization of product candidates, if a collaboration partner determines not to proceed with the future development of a program or product candidate, if a collaboration partner implements a clinical or regulatory strategy that ultimately does not enable the further development, approval or commercialization of the product candidate, or if a collaboration partner terminates its arrangement with us, we may not receive any future research funding or milestone, earnout, royalty or other contingent payments under such arrangement. In addition, our ability to monitor the achievement of clinical, regulatory and commercial milestones by our collaboration partners and enforce the payment of any corresponding fees is limited. If we do not receive the funding we expect under these agreements, the development of our product candidates could be delayed, we may need additional resources to develop such product candidates, and our research and development efforts and potential to generate revenue may be limited and our business, operating results, prospects and financial condition could be materially and adversely impacted.
We cannot assure investors that we will be able to maintain or expand our existing collaboration partners or that our City Platform will achieve adequate market acceptance among new collaboration partners. Any failure to increase penetration in our existing markets or new markets would adversely affect our ability to improve our operating results from our collaboration, partnership and licensing strategy.
All of the risks relating to product development, regulatory approval and commercialization described in this prospectus apply to the activities of our collaboration partners. If we and our
collaboration partners do not receive regulatory approval for a sufficient number of product candidates, we may not be able to sustain our business model.
We may seek to establish additional partnership, collaboration and licensing arrangements and, if we are not able to establish them on commercially reasonable terms, we may have to alter our development and commercialization plans. Certain of our partnership, collaboration and licensing arrangement may restrict our ability to develop certain products.
Our development programs and the potential commercialization of our product candidates will require substantial additional cash to fund expenses. For some of our product candidates, we may decide to collaborate with pharmaceutical and biotechnology companies for the development and potential commercialization of those product candidates. For example, we expect to seek a collaboration partner for our most advanced product candidate, CITY-FXI, prior to initiating Phase 3 development.
We face significant competition in seeking appropriate collaboration partners. Whether we reach a definitive agreement for any additional partnership, collaboration and licensing arrangements will depend, among other things, upon our assessment of the collaboration partner’s resources and expertise, the terms and conditions of the proposed partnership, collaboration or licensing arrangement and the proposed collaboration partner’s evaluation of a number of factors. Those factors may include the design or results of clinical trials, the likelihood of approval by Regulatory Authorities, the potential market for the subject product candidate, the costs and complexities of manufacturing and delivering such product candidate to trial participants, the potential of competing drugs, the existence of uncertainty with respect to our ownership of technology, which can exist if there is a challenge to such ownership without regard to the merits of the challenge, and industry and market conditions generally. The collaboration partner may also consider alternative product candidates or technologies for similar indications that may be available to collaborate on and whether such a collaboration could be more attractive than the one with us for our product candidate. The terms of any additional partnership, collaboration and licensing arrangements or other arrangements that we may establish may not be favorable to us.
Our existing collaboration and licensing agreements, and any agreements we may enter into in the future, may contain provisions that limit our operational flexibility, including field-of-use limitations, territory restrictions, or exclusivity provisions that preclude us from independently exploiting, or entering into arrangements with third parties with respect to, certain product candidates or technologies in defined areas. Any such restrictions could limit our ability to maximize the value of our pipeline and platform or to pursue new strategic opportunities. In addition, disputes with existing partners regarding the scope of any such restrictions could result in litigation or arbitration, divert management attention, and adversely affect our development and commercialization activities.
We may not be able to negotiate additional partnership, collaboration and licensing arrangements on a timely basis, on favorable terms or at all. Strategic alliances are complex and time-consuming to negotiate and document. If we are unable to negotiate and enter into new partnership, collaboration and licensing arrangements, we may have to curtail the development of the product candidate for which we are seeking to collaborate, reduce or delay its development program or one or more of our other development programs, delay its potential commercialization or reduce the scope of any sales or marketing activities, or increase our expenditures and undertake development or commercialization activities at our own expense. If we elect to increase our expenditures to fund development or commercialization activities on our own, we may need to obtain additional capital, which may not be available to us on favorable terms or at all. If we do not have sufficient funds, we may not be able to further develop our product candidates or bring them to market and generate product revenue.
If any collaborator materially amends, terminates or fails to perform its obligations under agreements with us, the development and commercialization of our products or product candidates could be delayed or terminated.
Our dependence on collaborators for capabilities and funding means that our business could be adversely affected if any collaborator materially amends or terminates its collaboration agreement with us, in whole or in part, or fails to perform its obligations under that agreement. Our current or future collaborations, if any, may not be scientifically or commercially successful. Disputes may arise in the
future with respect to the ownership of rights to technology or products developed with our collaborators, which could have an adverse effect on our ability to develop and commercialize any affected product candidate. Our current collaborations allow, and we expect that any future collaborations will allow, either party to terminate the collaboration for a material breach by the other party.
In addition, under certain circumstances, our collaborators may have additional termination rights for convenience with respect to the collaboration as a whole or a particular program under the collaboration. If we were to lose a commercialization collaborator, we would have to attract a new collaborator (potentially on less favorable terms for us than we have with our existing collaborator) or develop expanded sales, distribution and marketing capabilities internally, which would require us to invest significant financial and management resources, or discontinue the program. Following termination, we may not regain all rights to independently develop or commercialize the relevant product candidates or technologies, which could further limit our ability to extract value from the affected program. Any such outcome could result in significant delays, increased costs, and material adverse effects on our business, financial condition, and results of operations.
In addition, if we have a dispute with a collaborator over the ownership of technology or other matters, or if a collaborator terminates its collaboration with us, for breach or otherwise, or determines not to pursue the research, development and/or commercialization of the affected product or product candidate, it could delay our development of product candidates, result in the need for additional company resources to develop the impacted product candidate(s), require us to expend time and resources to develop expanded sales and marketing capabilities on a more expedited timeline, make it more difficult for us to attract new collaborators and adversely affect how we are perceived in the business and financial communities.
Moreover, a collaborator, or in the event of a change in control of a collaborator or the assignment of a collaboration agreement to a third party, the successor entity or assignee could determine that it is in its interests to:
•
pursue alternative technologies or develop alternative products, either on its own or jointly with others, that may be competitive with the products on which it is collaborating with us or which could affect its commitment to its collaboration with us;
•
pursue higher-priority programs or change the focus of its development programs, which could affect the collaborator’s commitment to us; or
•
if it has commercial rights, choose to devote fewer resources to the marketing of our products, if any receive regulatory approval, than it does for products developed or commercialized outside of our collaboration.
If any of these occur, the development and commercialization of one or more products or product candidates could be delayed, curtailed or terminated because we may not have sufficient financial resources or capabilities to continue such development and commercialization on our own.
We have no sales, distribution or marketing experience, and may invest significant financial and management resources to establish these capabilities. If we are unable to establish such capabilities or enter into agreements with third parties to market and sell our future products, if approved, we may be unable to generate any revenues.
Given our stage of development as a company, we have no sales, distribution or marketing experience. To successfully commercialize any products that may result from our programs, we will need to develop sales and marketing capabilities in the United States, Europe and other regions, either on our own or with others. These efforts will require substantial additional resources, some or all of which may be incurred in advance of any approval of these product candidates. Any failure or delay in the development of our or third parties’ internal sales, marketing and distribution capabilities would adversely impact the commercialization of our product candidates.
Factors that may inhibit our efforts to commercialize our product candidates on our own include:
•
inadequate funding;
•
our inability to recruit and retain an adequate number of effective sales and marketing personnel;
•
the inability of sales personnel to obtain access to or persuade an adequate number of physicians to prescribe any future products;
•
the lack of complementary products to be offered by sales personnel, which may put us at a competitive disadvantage compared to companies with more extensive product lines; and
•
unforeseen costs and expenses associated with creating an independent sales and marketing organization.
We may enter into partnership, collaboration and licensing arrangements with third parties to utilize their mature marketing and distribution capabilities, but we may be unable to enter into marketing agreements on favorable terms, if at all. If these third parties do not commit sufficient resources to commercialize our future products, if any, and we are unable to develop the necessary marketing capabilities on our own, we may be unable to generate sufficient product revenue to sustain our business. We may be competing with many companies that currently have extensive and well-funded marketing and sales operations. Without a significant internal team or the support of a third party to perform marketing and sales functions, we may be unable to compete successfully against these more established companies. Our future product revenue may be lower than if we directly marketed or sold our product candidates, if approved. In addition, any revenue we receive will depend in whole or in part upon the efforts of these third parties, which may not be successful and are generally not within our control. If we are not successful in commercializing any approved products, our future product revenue will suffer and we may incur significant additional losses.
If we do not establish sales and marketing capabilities successfully, either on our own or in collaboration with third-parties, we will not be successful in commercializing our product candidates.
Risks Related to Our Intellectual Property
Our success is largely based upon our intellectual property and proprietary technologies, and we may be unable to adequately protect and/or enforce our intellectual property.
Our success depends, in large part, on our ability to obtain and maintain patents, trademarks, trade secrets, know-how and other intellectual property rights and proprietary technology relating to our City Platform, our integrated RNAi platform, technologies and our product candidates, as well as our ability to successfully enforce our rights against third party infringers and/or defend our intellectual property against third party challenges or misappropriation. If we (or our licensees or licensors who may have the right to prosecute or enforce certain patents within our portfolio) fail to appropriately prosecute or are unable to obtain and maintain patent protection for our product candidates (or aspects thereof), our ability to develop, license and/or commercialize these product candidates may be adversely affected and we may not be able to prevent competitors from making, using, selling or importing competing products. This failure or inability to properly or adequately protect the intellectual property rights relating to these product candidates could have a material adverse effect on our business, financial condition, results of operations and/or prospects. We may lose a considerable amount of control over our intellectual property and may not receive anticipated revenues in strategic transactions, particularly where the consideration is contingent on the achievement of development or sales milestones.
RNAi therapeutic technology is not a new scientific field. We believe we are uniquely positioned to lead the next generation of RNAi therapeutic breakthroughs. Our differentiated City Platform is built on parallel innovation in two core areas: engineering the RNAi trigger, the double-stranded RNA that drives sequence-specific cleavage of target mRNA, and developing novel ligands for targeted extrahepatic delivery. We have applied for patents relating to our City Platform and some of our product candidates. The pending patent applications that we own or in-license in the United States and in key markets around the world, claim different aspects relating to our product candidates and potential product candidates and to the engineering, development, manufacture and commercialization of such product candidates including, but not limited to, compositions and methods of use.
The patent application process is subject to numerous risks and uncertainties, and there can be no assurance that we or our partners will be successful in protecting our product candidates by obtaining, maintaining, enforcing and defending patents. Patent applications are being processed by national patent offices around the world. There is uncertainty about which patents will issue, and, if they do, as to when, to whom, and with what claims. These risks and uncertainties include the following:
•
patent applications may not result in any patent being issued;
•
patents that may be issued may not include claims that cover a broad enough scope to prevent alternative solutions by competitors;
•
patents that may be issued may be challenged, invalidated, modified, revoked, circumvented, found to be unenforceable or otherwise may not provide adequate barriers to entry or any competitive advantage;
•
because of the extensive time required for development, testing and regulatory review of a product candidate, it is possible that before a potential product can be commercialized, any related patent may expire, or remain in existence for only a short period following commercialization, thereby reducing, or eliminating any advantage of the patent;
•
our competitors, many of which have substantially greater resources than we or our partners have, and many of which have made significant investments in competing technologies, may seek, or may already have sought or obtained, patents that will limit, interfere with or eliminate our ability to make, use and sell our product candidates;
•
there may be significant pressure on the United States government and other governmental bodies to limit the scope of patent protection or impose compulsory licensing of patent rights for disease treatments that prove successful as a matter of public policy regarding worldwide health concerns;
•
countries other than the United States may have less robust patent laws than those upheld by United States courts, allowing foreign competitors the ability to exploit these laws to create, develop, and market competing products using our technologies and patents; and
•
we may be involved in lawsuits and/or proceedings before government agencies, such as patent offices, to defend or enforce our patents or the patents we have rights to enforce, which could be expensive, time-consuming, distracting and/or unsuccessful.
In addition to patents, we also rely on proprietary trade secrets and know-how. Although we have taken steps to protect our unpatented proprietary trade secrets and know-how, including maintaining data security protocols and capabilities and entering into confidentiality agreements with third parties, and confidential information and assignment agreements with employees, consultants and advisors, there exists the potential that third parties may still somehow obtain this information or arrive at the same or similar information independently, which could reduce or eliminate our competitive advantages. Moreover, we may become subject to allegations that we directly or indirectly (through our consultants, advisors or independent contractors that we may engage to assist us in developing our product candidates) have wrongfully or inadvertently disclosed, acquired or used trade secrets or other proprietary information of third parties.
If our trade secrets are not adequately protected, our business, financial condition, results of operations and prospects could be adversely affected.
These risks are heightened due to our reliance on third parties, including third party CROs and CDMOs, for certain aspects of our business. The activities conducted by our third-party vendors require us to share our trade secrets with them, which increases the possibility that a competitor will discover them or that our trade secrets will be misappropriated or disclosed.
We may be forced to litigate to enforce or defend our intellectual property rights.
We may be forced to litigate to enforce or defend our intellectual property rights against infringement by competitors, and to protect our trade secrets and know-how against unauthorized use, but we may
not be able to detect or prevent, alone or with our licensors, infringement, misappropriation or other violations of our intellectual property rights. In so doing, we may place our intellectual property at risk of being invalidated, rendered unenforceable or limited or narrowed in scope such that we may no longer be able to adequately prevent the manufacture, sale or import of competitive products. In an infringement proceeding, a court may decide that a patent we own or a patent we may license in the future is invalid or unenforceable or may refuse to stop the other party from using the invention at issue. Grounds for an unenforceability assertion could be an allegation that someone connected with prosecution of the patent withheld relevant information from the United States Patent and Trademark Office (the “USPTO”) or made a misleading statement during prosecution. The outcome following legal assertions of invalidity and unenforceability is unpredictable. An adverse outcome in a litigation or proceeding involving our patents could limit our ability to assert our patents against those parties or other competitors and may curtail or preclude our ability to exclude third parties from making and selling similar or competitive products. Any of these occurrences could adversely affect our competitive position, business, financial condition, results of operations or prospects. Similarly, if we assert trademark infringement claims, a court may determine that the marks we have asserted are invalid or unenforceable, or that the party against whom we have asserted trademark infringement has superior rights to the marks in question. In this case, we could ultimately be forced to cease use of such trademarks.
Even if we establish infringement, the court may decide not to grant an injunction against further infringing activity and instead award only monetary damages, which may or may not be an adequate remedy. Moreover, an adverse result in any litigation or other proceedings before government agencies such as the USPTO, may place pending applications at risk of non-issuance or limitations in scope. Further, derivation proceedings, ex parte reexamination, inter partes review, post-grant review and opposition proceedings provoked by third parties or brought by the USPTO or any foreign patent authority may be used to challenge the inventorship, ownership, claim scope or validity of our patents. Additionally, because of the substantial amount of discovery typically required in connection with intellectual property litigation, there is a risk that some of our confidential and proprietary information, trade secrets or know-how could be compromised by disclosure during this type of litigation. In addition, there could be public announcements of the results of hearings, motions or other interim proceedings or developments, and if securities analysts or investors perceive these results to be negative, it could have a substantial adverse effect on the value of the company. Such litigation or proceedings could substantially increase our operating losses. reduce the resources available for development activities or any future sales, marketing or distribution activities and distract our personnel from their normal responsibilities. We may not have sufficient financial or other resources to conduct such litigation or proceedings adequately. Some of our competitors may be able to sustain the costs of such litigation or proceedings more effectively than we can because of their greater financial resources and more mature and developed intellectual property portfolios. Uncertainties resulting from the initiation and continuation of patent litigation or other proceedings could have a material adverse effect on our ability to compete in the marketplace.
The United States government and/or government agencies have provided funding or other assistance in connection with the development of the intellectual property rights owned by or licensed to us and if we enter into future arrangements involving government funding, and we make inventions as a result of such funding, our intellectual property rights to such discoveries may be subject to the applicable provisions of the Bayh Dole Act of 1980 (the “Bayh Dole Act”).
The United States government and/or government agencies have provided funding or other assistance in connection with the development of the intellectual property rights licensed to us, and if we enter into future arrangements involving government funding, and we make inventions as a result of such funding, our intellectual property rights to such discoveries may be subject to the applicable provisions of the Bayh Dole Act. To the extent any of our current and future intellectual property is generated through the use of United States government funding, the provisions of the Bayh Dole Act may similarly apply. If we enter into future arrangements involving government funding, any exercise by the government of certain rights could harm our competitive position, business, financial condition, results of operations and prospects.
United States government rights in certain inventions developed under a government-funded program include a non-exclusive, non-transferable, irrevocable worldwide license to use inventions for governmental purposes. In addition, the United States government would have the right to require us to grant exclusive, partially-exclusive or non-exclusive licenses to any of these inventions to a third party if the government determines that: (i) adequate steps have not been taken to commercialize the invention; (ii) government action is necessary to meet public health or safety needs or (iii) government action is necessary to meet requirements for public use under federal regulations, which are referred to as “march-in” rights. The United States government will also have the right to take title to these inventions if we fail, or the applicable licensor fails, to disclose the invention to the government, elect title and file an application to register the intellectual property within specified time limits. In addition, the United States government may acquire title to these inventions in any country in which a patent application is not filed within specified time limits. Intellectual property generated under a government funded program is also subject to federal regulations such as march-in rights, certain reporting requirements, compliance with which may require us, or the applicable licensor, to expend substantial resources. In addition, the United States government requires that any products embodying the subject invention or produced through the use of the subject invention be manufactured substantially in the United States. The manufacturing preference requirement can be waived if the owner of the intellectual property can show that reasonable but unsuccessful efforts have been made to grant licenses on similar terms to potential licensees that would be likely to manufacture substantially in the United States or that under the circumstances domestic manufacture is not commercially feasible.
If we become involved in patent litigation or other proceedings related to a determination of rights, we could incur substantial costs and expenses, substantial liability for damages or be required to stop our product development and commercialization efforts.
Our commercial success depends in part on our ability and the ability of any of our future partners to develop, manufacture, market and sell our product candidates and use our proprietary technologies without infringing on the intellectual proprietary rights of third parties. There is a substantial amount of litigation and patent office proceedings, both within and outside the United States, involving patent and other intellectual property rights in the biotechnology, biopharmaceutical, pharmaceutical and high-tech industries, including patent infringement lawsuits, oppositions, ex parte reexaminations, post-grant review, inter partes review, interference proceedings, and derivation proceedings before the USPTO and corresponding foreign patent offices. Numerous United States and foreign pending patent applications, which are owned by third parties, exist in the fields in which we are pursuing product candidates.
We may be subject to patent infringement claims, which could result in substantial costs and liability and prevent us from commercializing our potential products. Third parties may assert that we are employing their proprietary technology without authorization. There may be third-party patents or patent applications, which purport to claim compositions, formulations, methods of manufacture or methods for treatment relating to our product candidates, their manufacture or use. Because patent applications in most countries remain confidential for a period of time after they are filed (commonly, 18 months), it is possible that there are unpublished patent applications that may later issue with claims that our product candidates may be alleged to infringe. Because patent applications can take many years to issue, there may be pending patent applications which do not currently seem relevant, but may later result in issued patents that our product candidates may be alleged to infringe. In addition, third parties may obtain patents in the future and then allege that our technologies infringe upon these patents. Additionally, under United States patent law, a patent owner may seek a reissue within two years of issuance of a patent to broaden the scope of that patent’s claims. As a result, patents that, at the time of issuance, do not appear relevant to our activities may later be broadened in a manner that could impact our business. We cannot assure that any of our or our licensors’ patent searches or analyses, including the identification of relevant patents, the scope of patent claims or the expiration of relevant patents, are or will be complete or thorough, nor can we be certain that we or our licensors have identified or will identify each and every third-party patent and pending patent application in the United States and abroad that is relevant to or necessary for the development, manufacture, and commercialization of our current and future products and product candidates in any jurisdiction. Our interpretation of the relevance or the scope of a patent or a pending patent application may be incorrect, which may negatively
impact our ability to market our products. We may incorrectly determine that our products or product candidates are not covered by a third-party patent or may incorrectly predict whether a third party’s pending patent application will issue with claims of relevant scope. Alternatively, we may incorrectly determine that the Hatch-Waxman Amendments (as defined below) are a defense for a safe harbor to infringement of a patent we consider relevant to the research or clinical development of our product candidates. Our determination of the expiration date of any patent in the United States or abroad that we consider relevant may be incorrect, and we may incorrectly conclude that a third-party patent is invalid and unenforceable or not infringed. Our failure to identify and correctly interpret relevant patents may negatively impact our ability to develop, manufacture and market our products and product candidates. If we fail to identify and correctly interpret relevant patents, we may be subject to infringement claims. As the number of competitors in the market grows and the number of patents issued in this area increases, the possibility of patent infringement claims escalates. Defense of infringement and other claims, regardless of their merit, would involve substantial litigation expense and would be a substantial diversion of employee resources from our business.
Third party claims of intellectual property infringement may prevent or delay our development and commercialization efforts. We work in a highly competitive field. Numerous U.S. and foreign-issued patents and pending patent applications owned by third parties exist in the fields in which we plan to commercialize our products, including thromboembolic disease, Stargardt disease, and anemia of chronic disease, for which we are developing our product candidates using our proprietary City Platform technologies, as well as retinal diseases and CNS diseases, for which we are developing product candidates in collaborations with partners. Some of these applications may be confidential for a period of time after they are filed (commonly, 18 months). As we gain greater visibility and market exposure as a public company, the risk increases that our programs and commercialization activities may give rise to claims of infringement of the patent rights of others. We cannot assure you that our product candidates and other proprietary technologies we develop will not infringe existing or future patents owned by third parties. We may not be aware of patents that have already been issued for which a third party, such as a competitor in the fields in which we are developing our product candidates, might assert as infringed by us. It is also possible that patents owned by third parties of which we are aware, but which we do not believe we infringe or that we believe we have valid defenses to any claims of patent infringement, could be found to be infringed by us. It is not unusual that corresponding patents issued in different countries have different scopes of coverage, such that in one country a third-party patent does not pose a material risk, but in another country, the corresponding third-party patent may pose a material risk to any of our current or future product candidates.
If a third party alleges that we infringe its intellectual property rights, we may face a number of issues, including, but not limited to:
•
infringement and other intellectual property allegations, which, regardless of merit, may be expensive and time-consuming to litigate and may divert our management’s attention and financial resources from our core business;
•
substantial damages for infringement, which we may have to pay if a court decides that the product candidate or technology at issue infringes on or violates the third party’s rights, and, if the court finds that the infringement was willful, we could be ordered to pay treble damages and the patent owner’s attorneys’ fees (and, in certain jurisdictions outside of the United States, we could be ordered to pay the patent owner’s attorneys’ fees even without such finding);
•
a court prohibiting us from developing, manufacturing, importing, marketing or selling our product candidates, or from using our proprietary technologies, unless the third party licenses its product rights to use, which it is not required to do;
•
even if a license is available from a third party, we may have to pay substantial royalties, upfront fees, milestones and other amounts and/or grant cross-licenses to intellectual property rights for our products; and
•
redesigning our product candidates or processes so they do not infringe, which may not be possible or may require substantial monetary expenditures and time.
Any of the foregoing could have a material adverse effect on our business, results of operations, financial condition and prospects.
Our ability to commercialize our product candidates in the United States and abroad may be adversely affected if we cannot successfully defend against infringement allegations or obtain a license on commercially reasonable terms to relevant third party patents that cover our product candidates. Even if we have a strong defense and/or believe that third party intellectual property allegations are without merit, there can be no assurance that a court would find in our favor on questions of infringement, validity, enforceability and/or priority. A court of competent jurisdiction could hold that these third party patents are valid and enforceable and have been infringed upon, which could materially and adversely affect our ability to commercialize our product candidates or technologies covered by the asserted third party patents. In order to successfully challenge the validity of any such United States patent in federal court, we would need to overcome a presumption of validity. As this burden is high, which requires us to present clear and convincing evidence as to the invalidity of any such United States patent claims, there is no assurance that a court of competent jurisdiction would invalidate the asserted claims of any such United States patent.
If we are found to infringe a third party’s intellectual property rights, and we are unsuccessful in demonstrating that any such patents are invalid or unenforceable, any license we are able to obtain may be non-exclusive, thereby giving our competitors and other third parties access to the same technologies licensed to us, and may require us to pay substantial licensing fees and/or make ongoing royalty payments. If we are unable to obtain a necessary license to a third-party patent on commercially reasonable terms, we may be unable to commercialize our product candidates or such commercialization efforts may be significantly delayed, which could in turn significantly harm our business. We also could be found liable for significant monetary damages, including treble damages and attorneys’ fees, if we are found to have willfully infringed on a patent or other intellectual property right.
Some of our competitors may be able to sustain the costs of complex patent litigation more effectively than we can because they have substantially greater resources. In addition, any uncertainties resulting from the initiation and continuation of any litigation could have a material adverse effect on our ability to raise the funds necessary to continue our operations or could otherwise have a material adverse effect on our business, results of operations, financial condition and prospects.
Moreover, in recent years, individuals and groups that are non-practicing entities, commonly referred to as “patent trolls,” have acquired patents and other intellectual property assets for the purpose of making claims of infringement in order to extract settlements. From time to time, we may receive demand letters, notices or “invitations to license,” or may be the subject of claims that our products and business operations infringe or violate the intellectual property rights of others.
We may not be successful in obtaining or maintaining necessary intellectual property rights to product components and manufacturing processes for our development pipeline.
At present, we have rights to certain intellectual property through licenses from third parties and under patent filings that we own, to develop our product candidates. Because our pipeline may involve additional product candidates that could require the use of proprietary rights held by third parties, the growth of our business could depend in part on our ability to acquire, in-license or use these proprietary rights. In addition, our product candidates may require specific pharmaceutical formulations or delivery technologies to work effectively and efficiently, and these rights may be held by others. We may be unable to acquire or in-license intellectual property rights that may be necessary to permit us to implement our City Platform technologies or develop, manufacture or use our product candidates. The licensing and acquisition of third-party intellectual property rights is a competitive area, and a number of more established companies are also pursuing strategies to license or acquire third party intellectual property rights that we may consider attractive. These established companies may have a competitive advantage over us due to their size, cash resources, and greater clinical development and commercialization capabilities. Further, we may be unable to negotiate a license within the specified time frame or under terms that are acceptable to us. If we are unable to do so, the third party may offer the intellectual property rights to other parties, potentially blocking our ability to pursue our product candidate and enabling our competitors to compete with our product candidate.
We work in highly competitive fields, in which numerous United States and foreign-issued patents and pending patent applications owned by third parties exist. Competitive products and/or competitive advanced clinical assets owned by third parties exist. We may not be able to effectively secure first-tier technologies when competing against other companies or investors.
In addition, companies that perceive us to be a competitor may be unwilling to assign or license rights to us. We also may be unable to license or acquire third party intellectual property rights on terms that would allow us to make an appropriate return on our investment, or at all. If we are unable to successfully obtain rights to required third party intellectual property rights, our business, financial condition, results of operations and prospects could suffer.
We are dependent on intellectual property that we have in-licensed from third parties, and our failure to comply with our obligations under these licenses could result in loss of rights that are critical to our business.
We have in-licensed from third parties intellectual property that is material to our development programs and product candidates. Our rights under these licenses are subject to our compliance with ongoing obligations, which may include the payment of fees, milestones, and royalties; the achievement of development diligence milestones; the filing and prosecution of licensed patents; and other obligations. If we fail to comply with any such obligations, our licensors may have the right to terminate our licenses. Loss of our exclusivity or termination of one or more of our in-licenses could materially impair our ability to develop and commercialize our product candidates, could allow competitors to access the same technology, and could have a material adverse effect on our business, financial condition, results of operations and prospects. We may also need to renegotiate the terms of our licenses in the future, and we cannot guarantee that any renegotiation would result in terms favorable to us.
Our rights to develop and commercialize our product candidates are, and in the future, may be subject to the terms and conditions of licenses granted to us by others.
We are dependent on patent rights, know-how and proprietary technology licensed or otherwise acquired from third parties, and we may also enter into additional agreements with third parties in the future. Our current license agreements with third parties impose, and may in the future impose additional diligence, development and commercialization timelines, milestone payments, royalties, indemnification, insurance, non-competes or other obligations on us. If we fail to comply with our obligations to our licensors, collaborators or other third parties, our counterparties may have the right to terminate or take other actions under these agreements. Termination of these agreements or reduction or elimination of our rights under these agreements may result in us having to negotiate new or reinstated agreements with less favorable terms, or cause us to lose our rights under these agreements, including our rights to important intellectual property or technology that are necessary for our business. In particular, we depend on our exclusive license agreement with OSIF, pursuant to which we in-license patent rights related to the originating technology underlying cityRNAs and certain other aspects of our technology platform. As described elsewhere in this prospectus, OSIF may terminate the OSIF License Agreement for cause under specified circumstances.
Our success will depend in part on the ability of our licensors to obtain, maintain and enforce patent protection for our licensed intellectual property. Further, certain patent filings relating to our product candidates may now or in the future be subject to step-in rights of certain of our licensors. We have limited control over certain of our licensors, and may in the future have limited control of our other licensors, prosecution activities or use or licensing of any other intellectual property that may be related to our in-licensed intellectual property. Our licensors may not successfully prosecute the patent applications we license. Even if patents issue from these patent applications, our licensors may fail to maintain these patents, may determine not to pursue litigation against other companies that are infringing these patents, or may pursue such litigation less aggressively than we would. If any of our licensors or licensees having rights to file, prosecute, maintain and defend our patent rights fail to conduct these activities for patents or patent applications covering any of our product candidates, our ability to develop and commercialize those product candidates may be adversely affected and we may not be able to prevent competitors or other third parties from making, using or selling competing products. In
addition, we may sublicense certain of our rights under various third-party licenses to our collaboration partners. Any impairment of these sublicensed rights could result in reduced revenues under our partnership, collaboration or licensing arrangement or result in termination of an agreement by one or more of our collaboration partners. In addition, intellectual property rights that we may in-license in the future may be sublicensed under intellectual property owned by third parties, in some cases through multiple tiers. The actions of our licensors may therefore affect our rights to use our sublicensed intellectual property, even if we are in compliance with all of the obligations under our license agreements. Should our licensors or any of the upstream licensors fail to comply with their obligations under the agreements pursuant to which they obtain the rights that are sublicensed to us, or should such agreements be terminated or amended, our ability to develop and commercialize our product candidates may be materially harmed.
We cannot be certain that such activities by our licensors have been or will be conducted in compliance with applicable laws and regulations or will result in valid and enforceable patents or other intellectual property rights. Pursuant to the terms of the license agreements with our licensors, such licensors may have the right to control enforcement of our licensed patents or defense of any allegations asserting the invalidity of such patents and, even if we are permitted to pursue such enforcement or defense, we cannot ensure the cooperation of our licensors or, in some cases, other necessary parties, such as any co-owners of patents or other intellectual property from which we have not yet obtained a license. We cannot be certain that our licensors will allocate sufficient resources or prioritize their or our enforcement of such patents or defense of such allegations to protect our interests in the licensed patents. Even if we are not a party to these legal actions, an adverse outcome could harm our business because it might prevent us from continuing to license intellectual property that we may need to operate our business. In addition, even when we have the right to control patent prosecution of licensed patents and patent applications, enforcement of licensed patents, or defense of allegations asserting the invalidity of those patents, we may still be adversely affected or prejudiced by actions or inactions of our licensors and their counsel that took place prior to or after assuming control.
Our current or future license agreements may not provide exclusive or sufficient rights to use such intellectual property and technology in all relevant fields of use and in all territories in which we may wish to develop or commercialize our product candidates in the future. Some licenses granted to us may be subject to certain preexisting rights held by the licensors or certain third parties. As a result, we may not be able to prevent third parties from developing and commercializing competitive products in certain territories or fields.
In the event that our third-party licensors or other counterparties determine that, in spite of our efforts, we have breached a license agreement or have failed to meet certain obligations thereunder, it may elect to terminate the applicable agreement or, in some cases, one or more licenses under such agreement or otherwise restrict our rights under the agreement. Such termination or restriction of rights could result in us losing the ability to develop and commercialize product candidates and technology covered by the licensed intellectual property. In the event of such termination, or if the underlying patent rights under a third party in-license or other agreement fail to provide the intended exclusivity, third parties may be able to seek regulatory approval of, and to market, products identical to ours and we may be required to cease the development and commercialization of our product candidates. Moreover, our licensors may own or control intellectual property that has not been licensed to us and, as a result, we may be subject to allegations, regardless of their merit, that we are infringing or otherwise violating a licensor’s rights. Any of these events could have a material adverse effect on our competitive position, business, financial conditions, results of operations and prospects.
In addition, the agreements under which we license or otherwise acquire intellectual property or technology from third parties are complex, and certain provisions in such agreements may be susceptible to multiple interpretations. The resolution of any contract interpretation disagreement that may arise could narrow what we believe to be the scope of our rights to the relevant patents, know-how and proprietary technology, or increase what we believe to be our financial or other obligations under the relevant agreement. Disputes may also arise between us and our licensors or other counterparties regarding intellectual property subject to a license agreement, including: the scope of rights granted under the agreement and other interpretation-related issues; whether and the extent to which our
technology and processes infringe on intellectual property of the licensor that is not subject to the agreement; our right to sublicense patent and other rights to third parties under collaborative development relationships; our diligence obligations with respect to the use of the licensed or otherwise acquired technology in relation to our development and commercialization of our product candidates, and what activities satisfy those diligence obligations; and the ownership of inventions and know-how resulting from the joint creation or use of intellectual property by our licensors and us and our collaboration partners.
If disputes over intellectual property that we have licensed prevent or impair our ability to maintain our current licensing arrangements on favorable terms, we may be unable to successfully develop and commercialize the affected product candidates.
We are generally also subject to all of the same risks with respect to protection of intellectual property that we license, as we are for intellectual property that we own, which are described below. If we or our licensors fail to adequately protect this intellectual property, our ability to commercialize products could suffer.
If we are unable to protect the confidentiality of our proprietary trade secrets or know-how, our business and competitive position would be harmed.
In addition to patent protection, we rely heavily upon proprietary know-how protection and data security protocols and capabilities, as well as non-disclosure agreements and invention assignment agreements with our employees, consultants and third parties, to protect our confidential and proprietary information, especially where we do not believe patent protection is appropriate or obtainable.
It is our policy to require our employees, corporate collaborators, outside scientific collaborators, CROs, CDMOs, service providers, consultants, advisors and other third parties to execute confidentiality agreements upon the commencement of employment, consulting or business relationships with us. These agreements generally provide that all confidential information concerning our business or financial affairs developed by or made known to an individual or entity during the course of that party’s relationship with us are to be kept confidential and not disclosed to third parties, except in certain specified circumstances. In the case of employees, the agreements also provide that all inventions conceived by the individual, and that are related to our current or planned business or research and development or made during normal working hours, on our premises or using our equipment or proprietary information, are our exclusive property. In the case of consultants and other third-party service providers, the agreements provide us with certain rights to all inventions arising from the services provided to us by those individuals or entities. However, we cannot assure that we have entered into agreements with each party that may have or have had access to our proprietary technologies and processes. Additionally, the assignment of intellectual property rights may not be self-executing, or assignment agreements may be breached, and we may be forced to bring claims against third parties, or defend allegations that they may bring against us, to determine the ownership of what we regard as our intellectual property. We may not be able to obtain adequate remedies for any breaches of such agreements. Ultimately, enforcing a claim that a party wrongfully or illegally disclosed or misappropriated trade secrets or know-how can be difficult, expensive and time consuming, and the outcome is unpredictable.
In addition to contractual measures, we try to protect the confidential nature of our proprietary information through other appropriate precautions, such as physical and technological security measures. However, trade secrets and know-how can be difficult to protect despite these precautions. Such measures may not, for example, in the case of misappropriation of trade secrets or know-how by an employee, former employee, or third party with authorized access, provide adequate protection for our proprietary information. Our security measures may not prevent an employee, former employee, or consultant from misappropriating our trade secrets or know-how and providing them to a competitor, and recourse we take against such misconduct may not provide an adequate remedy to protect our interests fully. Enforcing a claim that a party wrongfully or illegally disclosed or misappropriated trade secrets or know-how can be difficult, expensive and time-consuming, and the outcome is unpredictable. In addition, trade secrets and know-how may be independently developed by others in a manner that could prevent legal recourse by us. If any of our confidential or proprietary information, such as our
trade secrets or know-how, were to be disclosed or misappropriated, or if any such information were independently developed by a competitor, our competitive position could be harmed.
Former employees may obtain employment with companies or academic institutions that could be considered competitive with us and are operating their business in areas that are similar to ours, including in their business model, product design efforts, product development or formulation technology. This competition may be limited by contractual provisions which may or may not be enforceable by us in the Commonwealth of Massachusetts or other jurisdictions. In addition, we may not be aware of such competitive employment arrangements until after our trade secrets or know-how has been disclosed to potentially competitive companies.
If we choose to go to court to stop a third party from using any of our trade secrets or know-how, we may incur substantial costs. In addition, courts inside and outside the United States are sometimes less willing or unwilling to protect trade secrets or know-how. Even if we are successful, these types of lawsuits may consume, in addition to substantial costs, significant amounts of our time and other resources. We may also need to share our proprietary know-how with current or future partners, collaborators, contractors and others located in countries at heightened risk of theft of trade secrets, including through direct intrusion by private parties or foreign actors, and those affiliated with or controlled by state actors. As a result, we may encounter significant problems in protecting and defending our intellectual property both in the United States and abroad. Any of the foregoing could have a material adverse effect on our business, financial condition, results of operations and prospects.
We may be subject to allegations that our employees, consultants or independent contractors have wrongfully used or disclosed confidential information of third parties or that our employees have wrongfully used or disclosed alleged trade secrets of their former employers.
As is common in the biotechnology industry, we employ individuals, including certain of our employees, who are or were previously employed at academic institutions or other biotechnology companies, including our competitors or potential competitors. For example, many of our co-founders and employees have held various positions in academia and in the RNAi industry. Although we seek to ensure that our employees, consultants and independent contractors do not use the proprietary information or know-how of others in their work for us, we may be subject to allegations that we, or our employees, consultants or independent contractors, have inadvertently or otherwise used or disclosed intellectual property, including trade secrets, know-how or other proprietary information, of any of our employees’ former employers or other third parties. Litigation may be necessary to defend against these allegations. If we fail in defending against any such allegations, in addition to paying monetary damages, we may lose valuable intellectual property rights or personnel, which could adversely impact our business. Even if we are successful in defending against such allegations, litigation could result in substantial costs and be a distraction to management and other employees.
We may be subject to allegations challenging the inventorship or ownership of our patents and other intellectual property.
We may be subject to allegations that former employees, collaborators or other third parties have an ownership interest in our patents or other intellectual property. Ownership disputes may arise, for example, from conflicting obligations of consultants or others who are involved in developing our product candidates. Litigation may be necessary to defend against these and other allegations challenging inventorship or ownership. If we fail in defending against any such allegations, in addition to paying monetary damages, we may lose valuable intellectual property rights, such as exclusive ownership of, or right to use, valuable intellectual property. Such an outcome could have a material adverse impact on our business. Even if we are successful in defending against such allegations, litigation could result in substantial costs and be a distraction to management and other employees.
In addition, while it is our policy to require our employees and contractors who may be involved in the conception or development of intellectual property to execute agreements assigning such intellectual property to us, we may be unsuccessful in executing such an agreement with each party who, in fact, conceives or develops intellectual property that we regard as our own. The assignment of intellectual property rights may not be self-executing, or the assignment agreements may be breached, and we may
be forced to bring claims against third parties, or defend claims that they may bring against us, to determine the ownership of what we regard as our intellectual property. Such claims could have a material adverse effect on our business, financial condition, results of operations and prospects.
Obtaining and maintaining our patent protection depends on compliance with various procedural, document submission, fee payment and other requirements imposed by governmental patent agencies, and our patent protection could be reduced or eliminated for non-compliance with these requirements.
Periodic maintenance fees, renewal fees, annuity fees and various other governmental fees on patents or applications will be due to be paid to the USPTO and various non-United States patent agencies in several stages over the lifetime of the patents or applications. The USPTO and non-United States patent agencies also require compliance with a number of procedural, documentary, fee payment and other similar provisions during the patent application process. We employ reputable law firms and other professionals to help us comply, and in many cases, an inadvertent lapse can be cured by payment of a late fee or by other means in accordance with the applicable rules. However, there are situations in which non-compliance can result in abandonment or lapse of the patent or patent application, resulting in partial or complete loss of patent rights in the relevant jurisdiction. In such an event, our competitors might be able to enter the market and this circumstance would have a material adverse impact on our business.
In addition, public health pandemics, geopolitical instability, natural disasters, or similar events may impair our and our licensors’ ability to comply with these procedural, document submission, fee payment, and other requirements imposed by government patent agencies, which may materially and adversely affect our ability to obtain or maintain patent protection for our product candidates. There could also be delays at the USPTO caused by staffing cuts and other United States government actions as a result of the United States Department of Government Efficiency or other executive actions to reduce the size of the United States government.
The USPTO and various non-United States government agencies require compliance with certain foreign filing requirements during the patent application process. For example, in some countries, including the United States, China, India, and some European countries, a foreign filing license is required before certain patent applications are filed. The foreign filing license requirements vary by country and depend on various factors, including where the inventive activity occurred, citizenship status of the inventors, the residency of the inventors and the invention owner, the place of business for the invention owner and the nature of the subject matter to be disclosed (e.g., items related to national security or national defense). In some, but not all cases, for example in China and India, a foreign filing license cannot be obtained retroactively in accordance with the applicable rules. There are situations, however, in which non-compliance can result in abandonment of a pending patent application or can be grounds for revoking or invalidating an issued patent, resulting in the loss of patent rights in the relevant jurisdiction. In such an event, potential competitors might be able to enter the relevant markets with similar or identical products or technology, which could have a material adverse effect on our business, financial condition, results of operations and prospects. We may also be dependent on our licensors to take the necessary actions to comply with these requirements with respect to our licensed intellectual property.
Issued patents covering our product candidates could be found invalid or unenforceable if challenged in court.
If we or one of our collaboration partners were to initiate legal proceedings against a third party to enforce a patent covering one of our product candidates, the defendant could counterclaim that the patent covering our product candidate is invalid and/or unenforceable. In patent litigation in the United States, counterclaims alleging invalidity or unenforceability are commonplace. Grounds for a validity challenge could be an alleged failure to meet any of several statutory requirements, including patent subject matter eligibility, novelty, non-obviousness, written description and/or enablement. Grounds for an unenforceability assertion could be an allegation that someone connected with prosecution of the patent withheld relevant information from the USPTO, or made a misleading statement, during prosecution. Third parties may also raise similar allegations before administrative bodies in the United
States or abroad, even outside the context of litigation. Such mechanisms include ex parte reexamination, inter partes review, post-grant review, interference proceedings, derivation proceedings, and equivalent proceedings in foreign jurisdictions (e.g., opposition proceedings). Such proceedings could result in revocation or amendment of our patents in such a way that they no longer cover our product candidates. The outcome following legal assertions of invalidity and unenforceability is unpredictable. With respect to the validity question, for example, we cannot be certain that there is no invalidating prior art, of which we and the patent examiner were unaware during prosecution. If a defendant were to prevail on a legal assertion of invalidity and/or unenforceability of patent rights covering a product candidate, we would lose at least part, and perhaps all, of the patent protection on our product candidates. Such a loss of patent protection could have a material adverse impact on our business. There is also a risk that, even if the validity of such patents is upheld, the court will construe the patent’s claims narrowly or decide that we do not have the right to stop the other party from using the invention at issue on the grounds that our patent claims do not cover the invention, or decide that the other party’s use of our patented technology falls under the safe harbor to patent infringement under 35 U.S.C. § 271(e)(1).
If we do not obtain sufficient patent term for our product candidates, our business may be materially harmed.
Patents have a limited term, and third parties may be able to circumvent our patents by developing similar or alternative products and technologies in a non-infringing manner, or develop and commercialize products and technologies similar or identical to ours and compete directly against us after the expiration of our patent rights, if any, and our ability to successfully commercialize any product or technology would be materially adversely affected. The terms of individual patents depend upon the legal term for patents in the countries in which they are granted. In most countries, including the United States, if all maintenance fees are timely paid, the natural expiration of a patent is generally 20 years from the earliest non-provisional filing date in the applicable country. However, the actual protection afforded by a patent varies from country to country, and also depends upon many factors, including the type of patent, the scope of coverage, the availability of regulatory-related extensions, the availability of extensions for patent office delays during the examination process, the availability of legal remedies in a particular country and the validity and enforceability of the patent, and whether a portion of the patent term has been terminally disclaimed based on other patents. These factors may emerge and change over the course of time, and accordingly, a patent’s expiration date might change over time in unpredictable ways. Various extensions including patent term extension and patent term adjustment may be available, but the durations of such extensions, and the protections they afford, are limited in the United States and other countries and regions. Additional patent terms may be available through a patent term adjustment process in the United States, resulting from USPTO delays during prosecution. Although various extensions may be available, the life of a patent, and the protection it affords, is limited. Even if patents covering our product candidates are obtained, once the patent life has expired for a product candidate, we may be open to competition from generics or biosimilars.
Depending upon the timing, duration and specifics of FDA regulatory approval of our product candidates, one or more patents issued from United States patent applications that we or a future licensor file may be eligible for limited patent term restoration under the Drug Price Competition and Patent Term Restoration Act of 1984 (the “Hatch-Waxman Amendments”). The Hatch-Waxman Amendments permit a patent restoration term of up to five years as compensation for patent term lost during the FDA regulatory review process based on the first regulatory approval for a particular drug or biologic. A maximum of one patent may be extended per FDA-approved drug as compensation for the patent term lost during the FDA regulatory review process. A patent term extension cannot extend the remaining term of a patent beyond a total of 14 years from the date of drug approval, and only those claims covering such approved drug product, a method for using it or a method for manufacturing it may be extended. Patent term extension may also be available in certain foreign countries upon regulatory approval of our product candidates.
Despite the possibility of an extension, we may not be granted an extension in the United States or another jurisdiction because of, for example, failure to exercise due diligence during the testing phase or regulatory review process, failing to apply within applicable deadlines, failing to apply prior to expiration
of relevant patents or otherwise failing to satisfy applicable requirements. Moreover, the applicable time or the scope of patent protection afforded could be less than we request.
If we are unable to obtain patent term extension or restoration, or the foreign equivalent, or the term of any such extension is less than we request, our competitors or other third parties may obtain approval of competing drugs following our patent expiration, and our revenue could be reduced, possibly materially. Further, if this occurs, our competitors or other third parties may take advantage of our investment in development and trials by referencing our clinical and preclinical data and launch their drug earlier than might otherwise be the case. Any of the foregoing could materially harm our business, financial condition, results of operations and prospects.
We may enjoy only limited geographical protection with respect to certain patents and we may not be able to protect our intellectual property rights throughout the world.
Filing and prosecuting patent applications and defending patents covering our product candidates in all countries throughout the world would be prohibitively expensive. Competitors may use our technologies and innovations in jurisdictions where we have not obtained patent protection to develop their own products and, further, may export otherwise infringing products to territories where we have patent protection, but enforcement rights are not as strong as those in the United States or Europe. These products may compete with our product candidates, and our and our licensors’ future patents or other intellectual property rights may not be effective or sufficient to prevent them from competing.
In addition, we may decide to abandon national and regional patent applications before they are granted. The examination of each national or regional patent application is an independent proceeding. As a result, patent applications in the same family may issue as patents in some jurisdictions, such as in the United States, but may issue as patents with claims of different scope or may even be refused in other jurisdictions. Furthermore, the requirements for patentability differ in certain jurisdictions and countries. Some countries do not grant claims directed to methods of treatment or have additional restrictions on the scope of method of treatment claims compared to the United States Accordingly, depending on the country, the scope of patent protection may vary for the same product candidate.
While we intend to protect our intellectual property rights in our expected significant markets, we cannot ensure that we will be able to initiate or maintain protection efforts in all such markets. Additionally, the prosecution of patent applications in other jurisdictions is often a longer process and patents may be granted at a later date than in the United States, potentially delaying our ability to assert such patents against competitors. Accordingly, our efforts to protect our intellectual property rights in such countries may be inadequate, which may have an adverse effect on our ability to successfully commercialize our product candidates in all of our expected significant foreign markets. If we encounter difficulties in protecting, or are otherwise precluded from effectively protecting, the intellectual property rights important for our business in such jurisdictions, the value of these rights may be diminished, and we may face additional competition in those jurisdictions.
The laws of some jurisdictions do not protect intellectual property rights to the same extent as the laws or rules and regulations in the United States and Europe, and many companies have encountered significant difficulties in protecting and defending such rights in such jurisdictions. The legal systems of certain countries, particularly certain developing countries, do not favor the enforcement of patents, trade secrets and other intellectual property rights, which could make it difficult for us to stop the infringement of any patents we obtain or marketing of competing products in violation of our proprietary rights generally. Proceedings to enforce our patent rights in other jurisdictions, whether or not successful, could result in substantial costs and divert our efforts and attention from other aspects of our business, could put any patents we obtain at risk of being invalidated or interpreted narrowly and our patent applications at risk of not issuing as patents, and could provoke third parties to legal claims against us. We may not prevail in any lawsuits that we initiate, and the damages or other remedies awarded, if any, may not be commercially meaningful. Accordingly, our efforts to enforce our intellectual property rights around the world may be inadequate to obtain a significant commercial advantage from the intellectual property that we develop or license, and this could negatively impact our business.
Some countries also have compulsory licensing laws under which a patent owner may be compelled to grant licenses to third parties. In addition, some countries limit the enforceability of patents against government agencies or government contractors. In those countries, the patent owner may have limited remedies, which could materially diminish the value of such patents. If we are forced to grant a license to third parties with respect to any patents relevant to our business, our competitive position may be impaired.
In Europe, a new unitary patent system took effect on June 1, 2023, which may significantly impact European patents, including those granted before the introduction of the new system. Under the new system, applicants can, upon grant of a European patent, opt for that patent to become a unitary patent which will be subject to the jurisdiction of a new unitary patent court (“UPC”). During the first seven years of the UPC’s existence, patents granted before the implementation of the new system can be opted out of UPC jurisdiction, and validated as national patents in any one or more of the UPC countries. We may decide to opt out future European patents from the UPC, but doing so may preclude us from realizing the benefits of the UPC. Moreover, if we do not meet all of the formalities and requirements for opt-out under the UPC, our future European patents could remain under the jurisdiction of the UPC. Patents that are under the jurisdiction of the UPC may be challenged in a single UPC-based revocation proceeding that, if successful, could invalidate the patent in all countries that are signatories to the UPC. The UPC will provide our competitors with a new forum to centrally revoke our European patents, and allow for the possibility of a competitor to obtain pan-European injunction. Further, because the UPC is a new court system and there is no precedent for the court’s laws, there is increased uncertainty regarding the outcome of any patent litigation. We are unable to predict what impact the new patent regime may have on our ability to exclude competitors in the European market. In addition to changes in patent laws, geopolitical dynamics, such as Russia’s invasion of Ukraine, may also impact our ability to obtain and enforce patents in particular jurisdictions, such as the enforcement of patent rights in Russia. If we are unable to obtain and enforce patents as needed in particular markets, our ability to exclude competitors in those markets may be reduced.
Changes in patent law could diminish the value of patents in general, thereby impairing our ability to protect our products.
As is the case with other biotechnology companies, our success is heavily dependent on intellectual property, particularly patents. Obtaining, defending, maintaining and enforcing patents in the biotechnology industry involves both technological and legal complexity and is therefore costly, time consuming and inherently uncertain. Changes in either the patent laws or interpretation of the patent laws in the United States and in other major jurisdictions could increase the uncertainties and costs surrounding the prosecution of patent applications and the enforcement or defense of issued patents, and may diminish our ability to protect our inventions, obtain, maintain, enforce and protect our intellectual property rights and, more generally, could affect the value of our intellectual property or narrow the scope of our future owned and licensed patents.
In addition, the patent positions of companies in the development and commercialization of biopharmaceuticals are particularly uncertain. Recent rulings from the United States Supreme Court and the Court of Appeals for the Federal Circuit have narrowed the scope of patent protection available in specified circumstances and weakened the rights of patent owners in specified situations. In addition to increasing uncertainty with regard to our ability to obtain patents in the future, this combination of events has created uncertainty with respect to the value of patents once obtained. Depending on decisions by the United States Congress, the federal courts and the USPTO, the laws and regulations governing patents could change in unpredictable ways that would weaken our ability to obtain new patents or to enforce our existing patents and patents that we might obtain in the future. In addition, the United States Supreme Court has ruled on several patent cases in recent years, either narrowing the scope of patent protection available in certain circumstances or weakening the rights of patent owners in certain situations. In addition to increasing uncertainty with regard to our ability to obtain patents in the future, this combination of events has created uncertainty with respect to the validity and enforceability of issued patents. Depending on future actions by the United States Congress, federal courts, the USPTO and the relevant law-making bodies in other countries, the laws and regulations governing patents could change in unpredictable ways that would weaken our or our licensors’ ability to obtain new
patents and patents that we or our licensors might obtain in the future. We cannot predict how future decisions by the federal courts, the United States Congress or the USPTO may impact the value of our patents. Any similar adverse change in the patent laws of other jurisdictions could also adversely affect our business, financial condition, results of operations and prospects.
For our United States patent applications, which contain claims entitled to a priority date after March 16, 2013, there is a greater level of uncertainty due to the Leahy-Smith America Invents Act (the “Leahy-Smith Act”), which was signed into law on September 16, 2011. The Leahy-Smith Act included a number of significant changes to United States patent law. These included provisions that affect the way patent applications are prosecuted, redefine prior art and provide more efficient and cost-effective avenues for competitors to challenge the validity of patents. The USPTO has promulgated regulations and developed procedures to govern administration of the Leahy-Smith Act, and many of the substantive changes to patent law associated with the Leahy-Smith Act did not come into effect until March 16, 2013. The Leahy-Smith Act and its implementation could increase the uncertainties and costs surrounding the prosecution of our patent applications and the enforcement or defense of our issued patents, all of which could have a material adverse effect on our business, financial condition, results of operations and prospects.
An important change introduced by the Leahy-Smith Act is that, as of March 16, 2013, the United States transitioned to a “first-to-file” system for deciding which party should be granted a patent when two or more patent applications are filed by different parties claiming the same invention. This requires us to be cognizant of the time from invention to filing of a patent application. Furthermore, our ability to obtain and maintain valid and enforceable patents depends on whether the differences between our technology and the prior art allow our technology to be patentable over the prior art. Since patent applications in the United States and most other countries are confidential for a period of time after filing, we cannot be certain that we were the first either to: (i) file any patent application related to our product candidates or (ii) invent any of the inventions claimed in our patents or patent applications.
Among some of the other changes introduced by the Leahy-Smith Act are changes that limit where a patentee may file a patent infringement suit and new procedures providing opportunities for third parties to challenge any issued patent in the USPTO. These new post grant challenges include post-grant review and inter partes review proceedings before the Patent Trial and Appeal Board at the USPTO. Because of a lower evidentiary standard in USPTO proceedings compared to the evidentiary standard in United States federal court necessary to invalidate a patent claim, a third party could potentially provide evidence in a USPTO proceeding sufficient for the USPTO to hold a claim invalid even though the same evidence would be insufficient to invalidate the claim if first presented in a district court action. Accordingly, a third party may attempt to use the USPTO procedures to invalidate patent claims that would not have been invalidated if first challenged by the third party as a defendant in a district court action. However, recent changes at the USPTO have resulted in many fewer inter partes review proceedings being instituted, and the USPTO has proposed modifications to the rules of practice for implementing inter partes review proceedings and ex parte reexamination. The proposed modifications, if adopted, could impact the ability of third parties, including us, to challenge the validity of granted United States patents before the USPTO.
Geopolitical actions in the United States and in foreign countries could increase the uncertainties and costs surrounding the prosecution or maintenance of patent applications and the maintenance, enforcement or defense of issued patents. For example, the United States and foreign government actions related to Russia’s invasion of Ukraine resulted in Russia issuing Decree No. 299 that effectively nullifies the enforcement of Russian patents owned by entities and individuals in “unfriendly” countries, including the United States
Similarly, changes in patent law and regulations in other countries or jurisdictions or changes in the governmental bodies that enforce them or changes in how the relevant governmental authority enforces patent laws or regulations may weaken our ability to obtain new patents or to enforce patents that we have licensed or that we may obtain in the future.
Any trademarks we have obtained or may obtain may be infringed or otherwise violated or successfully challenged. If our trademarks and trade names are not adequately protected, including if we are unable to obtain or establish desired trademarks rights or trade names rights, then we may not be able to build brand name recognition in our markets of interest and our business may be adversely affected.
We expect to rely on trademarks as one means to distinguish our product candidates, if approved for marketing, from third party products. Once we select new trademarks and apply to register them, our trademark applications may not be approved. During trademark registration proceedings in the United States and foreign jurisdictions, we may receive rejections. We are given an opportunity to respond to those rejections, but we may not be able to overcome such rejections.
We have also not yet registered trademarks for any of our product candidates in any jurisdiction. Any trademark applications we file may be rejected and registered trademarks may not be obtained, maintained or enforced. If we do not successfully register our trademarks, we may encounter difficulty in enforcing, or be unable to enforce, our trademark rights against third parties, which could adversely affect our business and our ability to effectively compete in the marketplace.
In addition, any proprietary name we propose to use with any of our product candidates in the United States will need to be approved by the FDA, regardless of whether we have registered, or applied to register, the proposed proprietary name as a trademark. The FDA conducts a review of proposed proprietary names, including an evaluation of potential for confusion with other products’ proprietary names. If the FDA objects to any of our proposed proprietary product names, we may be required to expend significant additional resources in an effort to identify a suitable alternative proprietary name that would qualify for protection under applicable trademark laws, not infringe the existing rights of third parties and be acceptable to the FDA.
In addition, our unregistered trademarks or trade names may be challenged, infringed, circumvented, declared generic, or determined to be infringing on, misappropriating or violating other marks. In the USPTO and in comparable agencies in many foreign jurisdictions, third parties are given an opportunity to oppose pending trademark applications and to seek to cancel registered trademarks. Opposition or cancellation proceedings may be filed against our trademarks, and our trademark registrations may not survive such proceedings. In the event that our trademarks are successfully challenged, we could be forced to rebrand our product candidates, which could result in loss of brand recognition and could require us to devote resources to advertising and marketing new brands. At times, competitors may adopt trade names or trademarks similar to ours, thereby impeding our ability to build brand identity and possibly leading to market confusion.
Our competitors may also infringe or otherwise violate our trademarks and we may not have adequate resources to enforce our trademarks. We may not be able to protect our rights to our trademarks and trade names, which we need to build name recognition among potential collaborators or customers in our markets of interest. Any of the foregoing events may have a material adverse effect on our business.
Furthermore, in many countries, owning and maintaining a trademark registration may not provide an adequate defense against a subsequent infringement allegation asserted by the owner of a senior trademark. Over the long term, if we are unable to successfully register our trademarks and trade names and establish name recognition based on our trademarks and trade names, then we may not be able to compete effectively, and our business may be adversely affected. Our efforts to enforce or protect our proprietary rights related to trademarks, trade names, domain names or other intellectual property may be ineffective and could result in substantial costs and diversion of resources and could adversely impact our financial condition or results of operations.
Trademark rights are territorial, and it is possible that we will not seek or successfully obtain trademark rights in certain jurisdictions that are relevant markets for the business, impeding our ability to compete effectively and adversely affecting our business.
Intellectual property rights do not necessarily address all potential threats.
The degree of future protection afforded by our intellectual property rights is uncertain because intellectual property rights have limitations and may not adequately protect our business or permit it to maintain our competitive advantage. For example:
•
our product candidates, if approved, may eventually become commercially available in generic or biosimilar product forms;
•
others may be able to make similar molecules to our product candidates that are not covered by the claims of the patents that we license or own now or in the future;
•
we, or current or future licensors or collaborators, might not have been the first to file patent applications covering certain of our or their inventions, potentially resulting in the invalidation of such patents or refusal of such applications;
•
we, or current or future licensors or collaborators, might not have been the first to make the inventions covered by the issued patent or pending patent application that we may license or own now or in the future.
•
we, or current or future licensors or collaborators, may fail to meet our obligations to the United States government regarding any patents and patent applications funded by United States government grants, leading to the loss or unenforceability of patent rights;
•
others may independently develop similar or alternative technologies or duplicate any of our technologies without infringing on our owned or licensed intellectual property rights;
•
it is possible that our pending patent applications or those that we may own or license in the future will not lead to issued patents;
•
it is possible that there are prior public disclosures that could invalidate our patents;
•
it is possible that there are unpublished patent applications that may later issue with claims covering our product candidates or technology similar to ours;
•
it is possible that our patents or patent applications omit individual(s) that should be listed as inventor(s) or include individual(s) that should not be listed as inventor(s), which may cause these patents or patents issuing from these patent applications to be held invalid or unenforceable or result in a change in ownership;
•
issued patents to which we hold rights may be held invalid, unenforceable or narrowed in scope, including as a result of legal challenges;
•
the claims of our issued patents or patent applications, if and when issued, may not cover our product candidates or narrowly cover them in such a way that competitors may be able to design around to avoid infringement allegations;
•
the laws of foreign countries may not protect our proprietary rights or the proprietary rights of current or future licensors or collaborators to the same extent as the laws of the United States;
•
the inventors of our patents or patent applications may become involved with competitors, develop products or processes that are similar to or alternative to those claimed in our patent filings or become hostile to our patents or patent applications on which they are named as inventors;
•
our competitors might conduct research and development activities in countries where we do not have patent rights and then use the information learned from such activities to develop competitive products for sale in our major commercial markets;
•
we have engaged in scientific collaborations in the past and we intend to continue to do so in the future, and our collaborators may develop adjacent or competing products that are outside the scope of our patents;
•
we may not develop additional proprietary technologies that are patentable;
•
the product candidates we develop may be covered by third party patents or other intellectual property rights;
•
the patents of others may prohibit or otherwise harm our ability to conduct our business; or
•
we may choose not to file a patent in order to maintain certain know-how, and a third party may subsequently commercialize the technology and/or file a patent covering such intellectual property.
Should any of these events occur, they could have a material adverse effect on our business, financial condition, results of operations and prospects.
Risks Related to Our Business Operations, Employee Matters and Industry
Our future success depends on our ability to retain employees, consultants and advisors and to attract, retain and motivate qualified personnel.
Our ability to compete in the highly competitive biotechnology industry depends upon our ability to attract and retain highly qualified managerial, scientific, technical and medical personnel. We are highly dependent upon members of our management, as well as technology and scientific teams, many of whom have been instrumental for us and have substantial experience with developing therapies, identifying potential product candidates and building the technologies related to the development of our City Platform and our pipeline. Each of the members of our management team, and all of our employees, including key technical personnel, scientists and clinicians, are employed “at will,” meaning we (or each officer or employee) may terminate the employment relationship at any time. The loss of any of these persons’ services may adversely impact the achievement of our research, development, financing and commercialization objectives. We currently do not have “key person” insurance on any of our employees. Many of our employees, including members of our executive team, have been with us for several years, and have stock options or other long-term equity incentives which may become valuable and will be publicly tradable if we become a public company. We may not be able to retain these employees due to the competitive environment in the biotechnology industry, particularly in the greater Boston, Massachusetts region.
In addition, we rely on consultants, contractors and advisors, including scientific and clinical advisors, to assist us in formulating our research and development, regulatory approval and commercialization strategy. We also rely on third-party consultants and contractors to support our internal finance, accounting and control function. Our consultants and advisors may be employed by employers other than us and may have commitments under consulting or advisory contracts with other entities that may limit their availability to us. The loss of the services of one or more of our current employees or advisors might impede the achievement of our research, development, regulatory approval and commercialization objectives. In addition, we have flexibly added capability and capacity through the use of contractors. We may not be able to retain the services of such personnel, which might result in delays in the operation of our business.
Recruiting and retaining other qualified employees, consultants and advisors for our business, including scientific and technical personnel, also will be critical to our success. Competition for skilled personnel, including in research, clinical operations, regulatory affairs, therapeutic area management and manufacturing, is intense and the turnover rate can be high. We may not be able to attract and retain personnel on favorable terms given the competition among numerous biotechnology companies and academic institutions for individuals with similar skill sets. In addition, adverse publicity, failure to succeed in preclinical or clinical trials or applications for marketing approval may make it more challenging to recruit and retain qualified personnel. The inability to recruit, or loss of services of certain executives, employees, consultants or advisors, may impede the progress of our research, development and commercialization objectives and have a material adverse impact on our business, financial condition, results of operations and prospects.
We expect to expand our organization, and as a result, we may encounter difficulties in managing our growth, which could disrupt our operations.
As we advance our research and development programs, we may need to further increase the number of our employees and the scope of our operations, particularly in the areas of clinical
development, biology, chemistry, manufacturing, general and administrative matters related to being a public company, regulatory affairs and, if any of our product candidates receives marketing approval, sales, marketing and distribution. To manage our anticipated future growth, we must:
•
expand our general and administrative functions;
•
identify, recruit, integrate, maintain and motivate additional qualified personnel;
•
manage our development efforts effectively, including the initiation and conduct of clinical trials for our product candidates;
•
establish and build a marketing and commercial organization; and
•
improve our operational, financial and management controls, reporting systems and procedures.
Our future financial performance and our ability to develop, manufacture and commercialize our product candidates, if approved, will depend, in part, on our ability to effectively manage any future growth, and our management may also have to divert financial and other resources, and a disproportionate amount of its attention away from day-to-day activities, to managing these growth activities.
If we are not able to effectively expand our organization by hiring new employees and expanding our groups of consultants and contractors, we may not be able to successfully implement the tasks necessary to further develop and commercialize our product candidates and, accordingly, may not achieve our research, development and commercialization goals.
Our employees, principal investigators and consultants may engage in misconduct or other improper activities, including non-compliance with regulatory standards and requirements and insider trading.
We are exposed to the risk of fraud or other misconduct by our employees, principal investigators and consultants. Misconduct by these parties could include intentional failures to comply with FDA regulations or the regulations applicable in the EU and other jurisdictions, provide accurate information to Regulatory Authorities, comply with healthcare fraud and abuse laws and regulations in the United States and abroad, report financial information or data accurately or disclose unauthorized activities to us. Such misconduct also could involve the improper use of information obtained in the course of clinical trials or interactions with Regulatory Authorities, which could result in regulatory sanctions and cause serious harm to our reputation. Sales, marketing and business arrangements in the healthcare industry are subject to extensive laws and regulations intended to prevent fraud, misconduct, kickbacks, self-dealing and other abusive practices. These laws and regulations restrict or prohibit a wide range of pricing, discounting, marketing and promotion, sales commission, customer incentive programs and other business arrangements. It is not always possible to identify and deter employee misconduct, and the precautions we take to detect and prevent this activity may not be effective in controlling unknown or unmanaged risks or losses or in protecting us from government investigations or other actions or lawsuits stemming from a failure to comply with these laws or regulations. If any such actions are instituted against us, and we are not successful in defending ourselves or asserting our rights, those actions could have a significant impact on our business, financial condition, results of operations and prospects, including the imposition of significant fines or other sanctions.
Clinical trial and product liability lawsuits against us could divert our resources and could cause us to incur substantial liabilities and to limit commercialization of any products that we may develop.
We face an inherent risk of clinical trial and product liability exposure related to the testing of our product candidates in clinical trials, and we will face an even greater risk if we commercially sell any products that we develop, if approved. While we currently have no products that have been approved for commercial sale, the ongoing, planned, and future use of product candidates by us in clinical trials, and the sale of any approved products in the future, may expose us to liability claims. These claims might be made by patients that use the product, healthcare providers, pharmaceutical companies, or others
selling such products. If we cannot successfully defend ourselves against claims that our product candidates or products caused injuries, we will incur substantial liabilities. Regardless of merit or eventual outcome, liability claims may result in:
•
decreased demand for any product candidates or products that we may develop;
•
termination of clinical trials;
•
injury to our reputation and significant negative media attention;
•
withdrawal of clinical trial participants;
•
significant costs to defend any related litigation;
•
substantial monetary awards to trial participants or patients;
•
loss of revenue;
•
reduced resources of our management to pursue our business strategy; and
•
the inability to commercialize any products that we may develop.
Although we currently hold clinical trial liability insurance, we will need to maintain such insurance coverage as we expand our clinical trials or if we commence commercialization of our product candidates. Insurance coverage is increasingly expensive. We may not be able to obtain and maintain insurance coverage at a reasonable cost or in an amount adequate to satisfy any liability that may arise. If a successful clinical trial or product liability claim or series of claims is brought against us for uninsured liabilities or in excess of insured liabilities, our assets may not be sufficient to cover such claims and our business operations could be impaired.
If the estimates we make, or the assumptions on which we rely, in preparing our consolidated financial statements prove inaccurate, our actual results may vary from those reflected in our accruals.
Our consolidated financial statements have been prepared in accordance with generally accepted accounting principles in the United States (“GAAP”). The preparation of these consolidated financial statements requires us to make estimates and judgments that affect the reported amounts of our assets, liabilities, revenues and expenses, the amounts of charges accrued by us and related disclosure of contingent assets and liabilities. We base our estimates on historical experience and on various other assumptions that we believe to be reasonable under the circumstances. We cannot assure you, however, that our estimates, or the assumptions underlying them, will be correct.
Changes in United States and international trade policies, particularly with respect to China, may adversely impact our business and operating results.
The United States government has recently made statements and taken certain actions that may lead to potential changes to United States and international trade policies, including imposing several rounds of tariffs and export control and sanctions restrictions affecting certain products manufactured in China. Both China and the United States have each imposed tariffs indicating the potential for further trade barriers, including the United States Commerce Department adding numerous Chinese entities to its Unverified List, which requires United States exporters to go through more procedures before exporting goods to such entities. It is unknown whether and to what extent new tariffs, export controls, or other new laws or regulations will be adopted, or the effect that any such actions would have on us or our industry. For example, in December 2025, the NDAA for Fiscal Year 2026 was enacted, which includes Section 851, commonly referred to as the “BIOSECURE Act.” The BIOSECURE Act restricts United States government agencies from procuring certain biotechnology equipment or services from, or entering into contracts with, entities that use biotechnology equipment or services from designated BCCs and from expending certain federal loan or grant funds for such equipment or services. While the BIOSECURE Act is primarily directed at United States government procurement and funding and has not yet been fully implemented through final regulations, there remains a continued policy interest in limiting United States companies’ relationships with biotechnology providers with relationships with foreign
adversaries. The potential downstream adverse impacts on entities having only commercial relationships with any impacted biotechnology providers is unknown but may include supply chain disruptions or delays. Sustained uncertainty about or further escalating trade and political tensions between the United States and China may prevent or hinder the export of materials or technical information among us, our CDMOs and other relevant third parties, such as pharmaceutical partners, or could result in trade or retaliatory restrictions that may hinder or potentially inhibit our ability to rely on CDMOs and other service providers that operate in China. Further, regulatory or legislative action taken by the United States to impose restrictions on transactions with China, like the restrictions described above, could have the potential to severely restrict the ability of companies like ours to contract with Chinese BCCs, which could have adverse effects on the development of our product candidates and our business operations.
Any unfavorable government policies on international trade, such as export controls, economic sanctions, capital controls or tariffs, may increase the cost of manufacturing our product candidates and platform materials, affect our ability to commercialize our product candidates, if approved, and adversely affect the import or export of raw materials and product candidates used in our research, preclinical studies and clinical trials, particularly to the extent we rely on third-party suppliers, manufacturers or other service providers located in China. If any new tariffs, export controls, sanctions, legislation or regulations are implemented, or if existing trade agreements are renegotiated or, in particular, if either the United States or Chinese government takes retaliatory trade actions due to recent trade tensions, such changes could have an adverse effect on our business, financial condition and results of operations.
Risks Related to Ownership of Our Common Stock and This Offering
There has been no prior public market for our common stock, and an active trading market for our common stock may not develop.
Prior to this offering, there was no public trading market for our common stock. If a market for our common stock does not develop or is not sustained, it may be difficult for you to sell your shares of our common stock at an attractive price or at all. We cannot predict the prices at which our common stock will trade. It is possible that in one or more future periods our results of operations may be below the expectations of public market analysts and investors, and, as a result of these and other factors, the price of our common stock may fall. An inactive market may also impair our ability to raise capital by selling our common stock and our ability to acquire other companies, products, or technologies by using our common stock as consideration.
The price of our common stock may be volatile and fluctuate substantially, which could result in substantial losses for purchasers of our common stock in this offering.
Our stock price is likely to be volatile. The stock market in general, and the market for biotechnology companies in particular, has experienced extreme volatility that has often been unrelated to the operating performance of particular companies. As a result of this volatility, you may not be able to sell your common stock at or above the initial public offering price. The market price for our common stock may be influenced by many factors, including: the commencement, enrollment, completion or results of preclinical and clinical trials of our product candidates or those of our competitors; the success of competitive products or technologies; commencement or termination of partnership, collaboration and licensing arrangements; regulatory or legal developments in the United States and other countries; developments or disputes concerning patent applications, issued patents or other proprietary rights; significant lawsuits, including patent or stockholder litigation; the recruitment or departure of key personnel; the level of expenses related to any of our product candidates or clinical development programs; the results of our efforts to discover, develop, acquire or in-license additional product candidates; actual or anticipated changes in estimates as to financial results, development timelines or recommendations by securities analysts; variations in our financial results or those of companies that are perceived to be similar to us; changes in the structure of healthcare payment systems; market conditions in the biotechnology and high-tech sectors, including high interest rates and borrowing costs; general economic, industry and market conditions; and the numerous product candidates in our
pipeline, the development of which could each generate news or significant AEs that could impact financial results or recommendations by securities analysts.
If our quarterly or annual results fall below the expectations of investors or securities analysts, the price of our common stock could decline substantially. Furthermore, any quarterly or annual fluctuations in our results may, in turn, cause the price of our stock to fluctuate substantially. We believe that period-to-period comparisons of our results are not necessarily meaningful and should not be relied upon as an indication of our future performance.
In the past, following periods of volatility in the market price of a company’s securities, securities class-action litigation often has been instituted against that company. Such litigation, if instituted against us, could cause us to incur substantial costs to defend such allegations and divert management’s attention and resources, which could seriously harm our business, financial condition, results of operations and prospects.
We may not be able to satisfy listing requirements of The Nasdaq Global Market (“Nasdaq”) or obtain or maintain a listing of our common stock on Nasdaq.
If, after listing, we fail to satisfy Nasdaq’s continued listing requirements, such as the corporate governance requirements or the minimum closing bid price requirement. Nasdaq may take steps to delist our common stock. Such a delisting would likely have a negative effect on the price of our common stock and would impair your ability to sell or purchase our common stock when you wish to do so. In the event of a delisting, we can provide no assurance that any action taken by us to restore compliance with listing requirements would allow our common stock to become listed again, stabilize the market price or improve the liquidity of our common stock, prevent our common stock from dropping below the Nasdaq minimum bid price requirement or prevent future non-compliance with Nasdaq’s listing requirements.
Our quarterly and annual operating results may fluctuate in the future. As a result, we may fail to meet or exceed the expectations of research analysts or investors, which could cause our stock price to decline and negatively impact our financing or funding ability, as well as negatively impact our ability to exist as a standalone company.
Our financial condition and operating results have varied in the past and will continue to fluctuate from quarter-to-quarter and year-to-year in the future due to a variety of factors, many of which are beyond our control and may be difficult to predict, including:
•
the timing and cost of, and level of investment in our City Platform and our research, development and, if approved, commercialization activities relating to our product candidates, which may change from time to time;
•
the timing and status of enrollment for clinical trials;
•
the cost of manufacturing our product candidates, as well as building out our supply chain, which may vary depending on the quantity of production and the terms of our agreements with manufacturers;
•
expenditures that we may incur to acquire, develop or commercialize additional product candidates and technologies;
•
timing and amount of any milestone, royalty or other payments due under any current or future collaboration, license, or acquisition agreement;
•
future accounting pronouncements or changes in our accounting policies;
•
the timing and success or failure of preclinical studies and clinical trials for our product candidates or competing product candidates, or any other change in the competitive landscape of our industry, including consolidation among our competitors or partners;
•
the timing of receipt of approvals for our product candidates from Regulatory Authorities in the United States and internationally;
•
exchange rate and interest rate fluctuations;
•
coverage and reimbursement policies with respect to our product candidates, if approved, and potential future drugs that compete with our products; and
•
the level of demand for our product candidates, if approved, which may vary significantly over time.
The cumulative effects of these factors could result in large fluctuations and unpredictability in our quarterly and annual operating results. As a result, comparing our operating results on a period-to-period basis may not be meaningful. Investors should not rely on our past results as an indication of our future performance.
This variability and unpredictability could also result in our failing to meet the expectations of industry or financial analysts or investors for any period. If our future revenue or operating results fall below the expectations of analysts or investors or below any forecasts we may provide to the market, or if any forecasts we provide to the market are below the expectations of analysts or investors, the price of our common stock could decline substantially. Such a stock price decline could occur even when we have met any previously publicly stated revenue or earnings guidance we may provide.
Future sales of our common stock in the public market could cause our stock price to fall.
Our stock price could decline as a result of sales of a large number of shares of our common stock after this offering or the perception that these sales could occur. These sales, or the possibility that these sales may occur, also might make it more difficult for us to sell equity securities in the future at a time and at a price that we deem appropriate.
Upon the completion of this offering, 50,332,279 shares of our common stock will be outstanding (or 51,790,612 shares of common stock will be outstanding assuming exercise in full of the underwriters’ option to purchase additional shares), based on our shares outstanding as of June 30, 2026, including 1,964,421 shares of restricted stock. All shares of our common stock expected to be sold in this offering will be freely tradable without restriction or further registration under the Securities Act of 1933, as amended (the “Securities Act”), unless held by our “affiliates,” as that term is defined in Rule 144 under the Securities Act. The resale of the remaining 40,610,057 shares, or 80.7% of our outstanding shares after this offering, is currently prohibited or otherwise restricted, subject to certain limited exceptions as described in the section titled “Underwriting,” as a result of securities law provisions, market standoff agreements entered into by our stockholders with us or lock-up agreements entered into by our stockholders with the underwriters. However, subject to applicable securities law restrictions and excluding shares of our restricted stock that will remain unvested, these shares will be able to be sold in the public market beginning 180 days after the date of this prospectus. Shares of our unvested restricted stock subject to repurchase or forfeiture that were issued and outstanding as of the date of this prospectus will become available for sale immediately upon the vesting of such shares, as applicable, and the expiration of any applicable market stand-off or lock-up agreements. Shares issued upon the exercise of stock options pursuant to future awards that may be granted under our equity incentive plans or pursuant to future awards granted under those plans will become available for sale in the public market to the extent permitted by the provisions of applicable vesting schedules, any applicable market stand-off and lock-up agreements and Rule 144 and Rule 701 under the Securities Act. For more information see the section titled “Shares Eligible for Future Sale.”
Upon the completion of this offering, the holders of approximately 28,848,641 shares, or 57.3%, of our common stock, will have rights, subject to some conditions, to require us to file registration statements covering the sale of their shares or to include their shares in registration statements that we may file for ourselves or other stockholders. We also intend to register the offer and sale of all shares of our common stock that we may issue under our equity compensation plans. Once we register the offer and sale of shares for the holders of registration rights and shares to be issued under our equity incentive plans, they can be freely sold in the public market upon issuance, subject to the lock-up agreements described in the section titled “Underwriting.”
In addition, certain of our executive officers, directors and stockholders affiliated with our directors may enter into Rule 10b5-1 plans providing for sales of shares of our common stock from time to time. Under a Rule 10b5-1 plan, a broker executes trades pursuant to parameters established by the executive officer, director or affiliated stockholder when entering into the plan, without further direction from the executive officer, director or affiliated stockholder. A Rule 10b5-1 plan may be amended or terminated in some circumstances. Our executive officers, directors and stockholders affiliated with our directors also may buy or sell shares outside of a Rule 10b5-1 plan when they are not in possession of material, nonpublic information.
In the future, we may also issue additional shares of our common stock or other equity or debt securities convertible into common stock in connection with a financing, acquisition, litigation settlement, employee arrangements or otherwise. Any such issuance could result in substantial dilution to our existing stockholders and could cause our stock price to decline.
If you purchase our common stock in this offering, you will incur immediate and substantial dilution in the book value of your shares.
You will suffer immediate and substantial dilution in the pro forma as adjusted net tangible book value of our common stock if you purchase in this offering. Assuming an initial public offering price of $18.00 per share, the midpoint of the price range set forth on the cover page of this prospectus, after giving effect to this offering, purchasers of common stock in this offering will experience immediate dilution of $11.47 per share in pro forma as adjusted net tangible book value of our common shares. In addition, after giving effect to this offering, investors purchasing common stock in this offering will contribute 37.9% of the total amount invested by stockholders since inception but will only own 20.1% of the common stock outstanding. In the past, we issued options and other securities to acquire common stock at prices significantly below the initial public offering price. To the extent these outstanding securities are ultimately exercised, investors purchasing common stock in this offering will sustain further dilution. See the section titled “Dilution” appearing elsewhere in this prospectus for a more detailed description of the dilution to new investors in the offering.
Raising additional capital may cause dilution to our existing stockholders, restrict our operations or require us to relinquish rights to our technologies or product candidates.
We may seek additional capital through public or private equity or debt financings, government or other third-party grants, asset sales, royalty financings, partnership, collaboration and licensing arrangements, or a combination of these approaches. To the extent that we raise additional capital through the sale of stock or convertible or exchangeable debt securities, warrants or other similar equity securities, your ownership interest could be diluted and the terms may include liquidation or other preferences that adversely affect your rights as a stockholder. The incurrence of indebtedness would result in increased fixed payment obligations and could involve restrictive covenants, such as limitations on our ability to incur additional debt, limitations on our ability to acquire or license intellectual property rights and other operating restrictions that could adversely impact our ability to conduct our business. If we raise additional funds through collaborations and alliances and licensing arrangements with third parties or through asset sales, we may have to relinquish valuable rights to our technologies or product candidates or grant licenses on terms unfavorable to us.
If securities analysts do not publish research or reports about our business or if they publish negative evaluations of our stock, the price of our stock could decline.
The trading market for our common stock will rely, in part, on the research and reports that industry or financial analysts publish about us or our business. We do not currently have, and may never obtain, research coverage by industry or financial analysts. If no, or few, analysts commence coverage of us, the trading price of our stock may decrease. Even if we do obtain analyst coverage, if one or more of the analysts covering our business downgrade their evaluations of our stock, the price of our stock could decline. If one or more of these analysts cease to cover our stock, we could lose visibility in the market for our stock, which in turn could cause our stock price to decline.
Our principal stockholders and management own a significant percentage of our stock and will be able to exert significant control over matters subject to stockholder approval.
Our executive officers, directors, five percent stockholders and their affiliates beneficially own approximately 69.1% of our common stock and, upon closing of this offering, that same group will beneficially own approximately 53.0% of our outstanding common stock (assuming no exercise of the underwriters’ option to purchase additional shares and no exercise of outstanding options and without giving effect to (i) any potential purchases by such persons in this offering or (ii) issuance of options to be granted to certain of our employees and non-employee directors upon pricing of this offering). Therefore, even after this offering, these stockholders will have the ability to influence us through their ownership positions. For example, these stockholders, acting together, may be able to exert significant influence over matters such as elections of directors, amendments of our organizational documents, or approval of any merger, sale of assets or other major corporate transaction. This may prevent or discourage unsolicited acquisition proposals or offers for our common stock that you may believe are in your best interest as one of our stockholders.
Some of these persons or entities may have interests different than yours. For example, because many of these stockholders purchased their shares at prices substantially below the current market price of our common stock and have held their shares for a longer period, they may be more interested in selling our company to an acquirer than other investors, or they may want us to pursue strategies that deviate from the interests of other stockholders.
Participation in this offering by our existing stockholders and their affiliated entities may reduce the public float for our common stock.
To the extent certain of our existing stockholders and their affiliated entities participate in this offering, such purchases would reduce the non-affiliate public float of our shares, meaning the number of shares of our common stock that are not held by officers, directors and principal stockholders. A reduction in the public float could reduce the number of shares that are available to be traded at any given time, thereby adversely impacting the liquidity of our common stock and depressing the price at which you may be able to sell shares of common stock purchased in this offering.
We have broad discretion in the use of our cash and cash equivalents, including the net proceeds from this offering, and may not use these proceeds effectively, affecting our results of operations and causing a decline in our stock price.
Our management will have broad discretion in the application of our cash and cash equivalents, including the net proceeds from this offering, and could spend the proceeds in ways that do not improve our results of operations or enhance the value of our common stock. The failure by our management to apply these funds effectively could result in financial losses that could have a material adverse impact on our business, cause the price of our common stock to decline, and delay the development of our product candidates. Pending their use, we may invest our cash and cash equivalents, including the net proceeds from this offering, in a manner that does not produce income or that loses value. See the section titled “Use of Proceeds” appearing elsewhere in this prospectus.
Provisions in our amended and restated certificate of incorporation, which will become effective immediately prior to the closing of this offering, and amended and restated bylaws, which will become effective upon the effectiveness of the registration statement of which this prospectus forms a part, and Delaware law may have anti-takeover effects that could discourage an acquisition of us by others, even if an acquisition would be beneficial to our stockholders, and may prevent attempts by our stockholders to replace or remove our current management.
Our amended and restated certificate of incorporation, which will become effective immediately prior to the closing of this offering, and amended and restated bylaws, which will become effective upon the effectiveness of the registration statement of which this prospectus forms a part, and Delaware law contain provisions that may have the effect of discouraging, delaying or preventing a change in control of us or changes in our management that stockholders may consider favorable, including transactions in which
you might otherwise receive a premium for your shares. Our amended and restated certificate of incorporation and amended and restated bylaws include provisions that:
•
authorize “blank check” preferred stock, which could be issued by our board of directors without stockholder approval and may contain voting, liquidation, dividend and other rights superior to our common stock;
•
create a classified board of directors whose members serve staggered three-year terms;
•
specify that special meetings of our stockholders can be called only by our board of directors;
•
prohibit stockholder action by written consent;
•
establish an advance notice procedure for stockholder approvals to be brought before an annual meeting of our stockholders, including proposed nominations of persons for election to our board of directors;
•
provide that vacancies on our board of directors may be filled only by a majority of directors then in office, even though less than a quorum;
•
provide that our directors may be removed only for cause;
•
specify that no stockholder is permitted to cumulate votes at any election of directors;
•
expressly authorize our board of directors to make, alter, amend or repeal our amended and restated bylaws; and
•
require supermajority votes of the holders of our common stock to amend specified provisions of our amended and restated certificate of incorporation and amended and restated bylaws.
These provisions, alone or together, could delay or prevent hostile takeovers and changes in control or changes in our management. These provisions could also limit the price that investors might be willing to pay in the future for shares of our common stock, thereby depressing the market price of our common stock.
In addition, because we are incorporated in the State of Delaware, we are governed by the provisions of Section 203 of the Delaware General Corporation Law (“DGCL”), which prohibits a person who owns in excess of 15% of our outstanding voting stock from merging or combining with us for a period of three years after the date of the transaction in which the person acquired in excess of 15% of our outstanding voting stock, unless the merger or combination is approved in a prescribed manner.
Any provision of our amended and restated certificate of incorporation, amended and restated bylaws or Delaware law that has the effect of delaying or deterring a change in control could limit the opportunity for our stockholders to receive a premium for their shares of our common stock and could also affect the price that some investors are willing to pay for our common stock.
Because we do not anticipate paying any cash dividends on our capital stock in the foreseeable future, capital appreciation, if any, will be your sole source of gain.
We do not currently intend to declare or pay cash dividends on our capital stock. We currently intend to retain all of our future earnings, if any, to finance the growth and development of our business. In addition, the terms of any future debt agreements may preclude us from paying dividends. As a result, capital appreciation, if any, of our common stock will be your sole source of gain for the foreseeable future.
Our amended and restated bylaws which will become effective upon the effectiveness of this registration statement of which this prospectus forms a part designate certain courts as the sole and exclusive forum for certain types of actions and proceedings that may be initiated by our stockholders, which could limit our stockholders’ ability to obtain a favorable judicial forum for disputes with us or our directors, officers, or employees.
Our amended and restated bylaws which will become effective upon effectiveness of the registration statement of which this prospectus forms a part provide that, unless we consent in writing to an alternative
forum, the Court of Chancery of the State of Delaware will be the sole and exclusive forum for any state law claims for (i) any derivative action or proceeding brought on our behalf, (ii) any action asserting a claim of breach of, or a claim based on, fiduciary duty owed by any of our current or former directors, officers, and employees to us or our stockholders, (iii) any action asserting a claim arising pursuant to any provision of the DGCL, our amended and restated certificate of incorporation or our amended and restated bylaws (including the interpretation, validity or enforceability thereof) or (iv) any action asserting a claim that is governed by the internal affairs doctrine (Delaware Forum Provision). The Delaware Forum Provision will not apply to any causes of action arising under the Securities Act or the Exchange Act of 1934, as amended (the “Exchange Act”). Our amended and restated bylaws further provide that, unless we consent in writing to the selection of an alternative forum, the federal district courts of the United States shall be the sole and exclusive forum for resolving any complaint asserting a cause or causes of action arising under the Securities Act (“Federal Forum Provision”). In addition, our amended and restated bylaws provide that any person or entity purchasing or otherwise acquiring any interest in shares of our common stock is deemed to have notice of and consented to the foregoing provisions; provided, however, that stockholders cannot and will not be deemed to have waived our compliance with the federal securities laws and the rules and regulations thereunder.
The Delaware Forum Provision and the Federal Forum Provision in our amended and restated bylaws may impose additional litigation costs on stockholders in pursuing any such claims. Additionally, the forum selection clauses in our amended and restated bylaws may limit our stockholders’ ability to bring a claim in a judicial forum that they find favorable for disputes with us or our directors, officers or employees, which may discourage such lawsuits against us and our directors, officers and employees even though an action, if successful, might benefit our stockholders. In addition, while the Delaware Supreme Court and other state courts have upheld the validity of federal forum selection provisions purporting to require claims under the Securities Act be brought in federal court, there is uncertainty as to whether other courts will enforce our Federal Forum Provision. If the Federal Forum Provision is found to be unenforceable, we may incur additional costs associated with resolving such matters. The Federal Forum Provision may also impose additional litigation costs on stockholders who assert that the provision is not enforceable or invalid. The Court of Chancery of the State of Delaware and the federal district courts of the United States may also reach different judgments or results than would other courts, including courts where a stockholder considering an action may be located or would otherwise choose to bring the action, and such judgments may be more or less favorable to us than our stockholders.
General Risk Factors
We will incur increased costs as a result of operating as a public company, and our management will be required to devote substantial time to new compliance initiatives. We will be subject to financial reporting and other requirements for which our accounting and other management systems and resources may not be adequately prepared.
As a public company, and particularly after we are no longer an “emerging growth company,” we will incur significant legal, accounting and other expenses that we did not incur as a private company. In addition, the federal securities laws, including the Sarbanes-Oxley Act and rules subsequently implemented by the SEC and Nasdaq have imposed various requirements on public companies, including requirements to file annual, quarterly and event driven reports with respect to our business and financial condition, and to establish and maintain effective disclosure and financial controls and corporate governance practices. Our management and other personnel will need to devote a substantial amount of time to these compliance initiatives. Moreover, these rules and regulations will increase our legal and financial compliance costs and will make some activities more time-consuming and costly. For example, we expect that these rules and regulations may make it more difficult and more expensive for us to obtain director and officer liability insurance. We may not be able to produce reliable financial statements or file these financial statements as part of a periodic report in a timely manner with the SEC or comply with the Nasdaq listing requirements. In addition, we could make errors in our financial statements that could require us to restate our financial statements.
We are an “emerging growth company,” and the reduced disclosure requirements applicable to emerging growth companies may make our common stock less attractive to investors.
We are an “emerging growth company” (“EGC”), as defined in the JOBS Act. We will remain an EGC until the earlier of: (i) the last day of the fiscal year in which we have total annual gross revenues of $1.235 billion or more; (ii) the last day of the fiscal year following the fifth anniversary of the date of the completion of this offering; (iii) the date on which we have issued more than $1.0 billion in non-convertible debt during the previous three years; or (iv) the date on which we are deemed to be a large accelerated filer under the rules of the SEC, which means the first day of the year following the first year in which the market value of our common stock that is held by non-affiliates exceeds $700 million. For so long as we remain an EGC, we are permitted and intend to rely on exemptions from certain disclosure requirements that are applicable to other public companies that are not emerging growth companies. These exemptions include: not being required to comply with the auditor attestation requirements of Section 404 of the Sarbanes-Oxley Act (“Section 404”); not being required to comply with any requirement that may be adopted by the Public Company Accounting Oversight Board regarding mandatory audit firm rotation or a supplement to the auditor’s report providing additional information about the audit and the financial statements; reduced disclosure obligations regarding executive compensation; and exemptions from the requirements of holding a non-binding advisory vote on executive compensation and shareholder approval of any golden parachute payments not previously approved.
We may choose to take advantage of some, but not all, of the available exemptions. We have taken advantage of reduced reporting burdens in this prospectus. In particular, we have not included all of the executive compensation information that would be required if we were not an EGC. We cannot predict whether investors will find our common stock less attractive if we rely on certain or all of these exemptions. If some investors find our common stock less attractive, as a result, there may be a less active trading market for our common stock and our stock price may be more volatile.
In addition, the JOBS Act provides that an EGC may take advantage of an extended transition period for complying with new or revised accounting standards. This allows an EGC to delay the adoption of certain accounting standards until those standards would otherwise apply to private companies. We have elected to avail ourselves of this exemption and, therefore, while we are an EGC we will not be subject to new or revised accounting standards at the same time that they become applicable to other public companies that are not EGCs.
If we fail to establish and maintain proper and effective internal control over financial reporting, our operating results and our ability to operate our business could be harmed.
Ensuring that we have adequate internal financial and accounting controls and procedures in place so that we can produce accurate financial statements on a timely basis is a costly and time-consuming effort that needs to be re-evaluated frequently. Our internal control over financial reporting is a process designed to provide reasonable assurance regarding the reliability of financial reporting and the preparation of financial statements in accordance with generally accepted accounting principles. In connection with this offering, we intend to begin the process of documenting, reviewing and improving our internal controls and procedures for compliance with Section 404 of the Sarbanes-Oxley Act, which will require annual management assessment of the effectiveness of our internal control over financial reporting starting with our second filing of an Annual Report on Form 10-K.
Implementing any appropriate changes to our internal controls may distract our officers and employees, entail substantial costs to modify our existing processes and take significant time to complete. These changes may not, however, be effective in maintaining the adequacy of our internal controls, and any failure to maintain that adequacy or consequent inability to produce accurate financial statements on a timely basis could increase our operating costs and harm our business. In addition, investors’ perceptions that our internal controls are inadequate or that we are unable to produce accurate financial statements on a timely basis cause investors to lose confidence in the accuracy and completeness of our financial reports and could cause the market price of our common stock to decline significantly.
Our disclosure controls and procedures may not prevent or detect all errors or acts of fraud.
Upon the closing of this offering, we will become subject to the periodic reporting requirements of the Exchange Act. We designed our disclosure controls and procedures to reasonably assure that information we must disclose in reports we file or submit under the Exchange Act is accumulated and communicated to management, and recorded, processed, summarized and reported within the time periods specified in the rules and forms of the SEC. We believe that any disclosure controls and procedures or internal controls and procedures, no matter how well-conceived and operated, can provide only reasonable, not absolute, assurance that the objectives of the control system are met.
These inherent limitations include the facts that judgments in decision-making can be faulty and that breakdowns can occur because of simple error or mistake. Additionally, controls can be circumvented by the individual acts of some persons, by collusion of two or more people or by an unauthorized override of the controls. Accordingly, because of the inherent limitations in our control system, misstatements due to error or fraud may occur and not be detected.
Our future ability to utilize our NOL carryforwards and certain other tax attributes may be limited.
Since our inception, we have incurred losses and we may never achieve profitability. As of December 31, 2025, we had United States federal net operating loss (“NOL”) carryforwards of $47.1 million (which generally are not subject to expiration), state NOL carryforwards of $49.2 million (which begin to expire in various amounts in 2045), United States federal research credit carryforwards of $2.0 million (which expire at various dates through 2045), and state research credit carryforwards of $1.2 million (which expire at various dates through 2040). Our unused United States federal NOLs may be carried forward to offset a portion of future taxable income, if any, but the deductibility of such United States federal NOL carryforwards in a taxable year is limited to 80% of taxable income in such year. Additionally, we continue to generate business tax credits, including research and development tax credits, which generally may be carried forward to offset a portion of future taxable income, if any, subject to expiration of such credit carryforwards. Under Sections 382 and 383 of the Internal Revenue Code of 1986, as amended (the “Code”), if a corporation undergoes an “ownership change,” generally defined as one or more shareholders or groups of shareholders that own at least five percent of the corporation’s equity increasing their equity ownership in the aggregate by more than 50 percentage points (by value) over a three-year period, the corporation’s ability to use its pre-change NOLs and other pre-change tax attributes (such as research and development tax credits) to offset its post-change income or taxes may be limited. Similar rules may apply under state tax laws. Our prior equity offerings and other changes in our stock ownership have resulted in such ownership changes in the past. In addition, we may experience ownership changes in the future as a result of this offering or subsequent shifts in our stock ownership, some of which are outside of our control. As a result, if we earn net taxable income, our ability to use our pre-change NOLs or other pre-change tax attributes to offset United States federal taxable income or taxes may be subject to limitations, which could potentially result in increased future tax liability to us. There is a risk that due to changes under the tax law, regulatory changes or other unforeseen reasons, our existing NOLs or business tax credits could expire or otherwise be unavailable to offset future income tax liabilities. At the state level, there may also be periods during which the use of NOLs or business tax credits is suspended or otherwise limited, which could accelerate or permanently increase state taxes. For these reasons, we may not be able to realize a tax benefit from the use of our NOLs or tax credits, even if we attain profitability.
Changes in tax laws or in their implementation or interpretation may adversely affect our business and financial condition.
The rules dealing with United States federal, state and local income taxation are constantly under review by persons involved in the legislative process and by the Internal Revenue Service, the United States Treasury Department and other taxing authorities. Changes to tax laws (which changes may have retroactive application) could adversely affect our business and our financial condition. In recent years, many such changes have been made and changes are likely to continue to occur in the future. We cannot predict whether, when, in what form or with what effective dates, tax laws, regulations and rulings
may be enacted, promulgated or decided or whether they could increase our tax liability or require changes in the manner in which we operate in order to minimize increases in our tax liability.
If our information technology systems or our infrastructure or those of third parties with whom we work, or our data, are or were compromised, we could experience adverse consequences resulting from such compromise that negatively impact our business and operations and subject us to liability.
In the ordinary course of our business, we and the third parties with whom we work, collect, receive, store, process, generate, use, transfer, disclose, make accessible, protect, secure, dispose of, transmit, and share (collectively, process) proprietary, confidential, and sensitive data, including personal data (such as health-related data), intellectual property, trade secrets, and data related to our clinical trials (collectively, sensitive information). As a result, the confidentiality, integrity, and availability of our information technology systems and sensitive information, and those of the third parties with whom we work, are vulnerable to compromise or damage from a variety of threats, including cybersecurity attacks, malicious internet-based activity, online and offline fraud, and other similar activities. Such threats are prevalent and continue to rise, are increasingly difficult to detect, and come from a variety of sources, including traditional computer “hackers,” threat actors, “hacktivists,” organized criminal threat actors, personnel (such as through theft or misuse), sophisticated nation states, and nation-state-supported actors.
We have experienced significant growth in the complexity of our data and the software tools that our hardware infrastructure supports. As a result, we rely significantly upon information technology systems and infrastructure owned and maintained by us or by third-party providers to process our sensitive information and to operate our business. We also outsource elements of our operations to, and obtain products and services from, third parties and engage in collaborations for drug design with third parties, each of which has or could have access to our sensitive information.
We deploy and operate an array of technical and procedural controls designed to reduce the risks to our information technology systems, infrastructure and sensitive information and to maintain the availability, confidentiality and integrity of our sensitive information, and we expect to continue to incur significant costs related to such detection and prevention efforts. While we continue to make investments to detect, mitigate, contain, and remediate vulnerabilities in our information technology systems, including through hiring qualified information technology personnel, conducting periodic cyber security awareness trainings, improving our information technology infrastructure and related controls, performing systematic back-ups of our data and systems, and conducting regular testing of our systems, there can be no assurance that our efforts will be effective. Actions taken by us or the third parties with whom we work to detect, investigate, mitigate, contain, and remediate a security incident could result in outages, data losses, and disruptions of our business. Threat actors may also gain access to other networks and systems after a compromise of our networks and systems. For example, threat actors may use an initial compromise of one part of our environment to gain access to other parts of our environment or leverage a compromise of our networks or systems to gain access to the networks or systems of third parties with whom we work, such as through phishing or supply chain attacks.
Despite these efforts, we and the third parties with whom we work are subject to a variety of evolving threats, including but not limited to social-engineering attacks (including through deep fakes, which may be increasingly more difficult to identify as fake, and phishing attacks), malicious code (such as viruses and worms), malware (including as a result of advanced persistent threat intrusions), denial-of-service attacks, credential stuffing attacks, credential harvesting, personnel misconduct or error, ransomware attacks, supply-chain attacks, software bugs, server malfunctions, software or hardware failures, loss of data or other information technology assets, adware, telecommunications failures, earthquakes, fires, floods, attacks enhanced or facilitated by artificial intelligence (“AI”), and other similar threats.
In particular, severe ransomware attacks are becoming increasingly prevalent and can lead to significant interruptions in our operations, ability to provide our products or services, loss of sensitive information and income, reputational harm, and diversion of funds. Extortion payments may alleviate the
negative impact of a ransomware attack, but we may be unwilling or unable to make such payments due to, for example, applicable laws or regulations prohibiting such payments.
With respect to cyber-attacks, the techniques used by cyber criminals change frequently, may not be recognized until launched, and can originate from a wide variety of sources, including outside groups and individuals with a range of motives (including industrial espionage) and expertise, such as organized crime affiliates, terrorist organizations or hostile foreign governments or agencies. These risks may be heightened in connection with geopolitical events or in conjunction with military conflicts and defense activities. During times of war and other major conflicts, we, the third parties with whom we work, may be vulnerable to a heightened risk of these attacks, including retaliatory cyber-attacks, that could materially disrupt our systems and operations, supply chain, and ability to produce and distribute our goods and services.
The costs to detect, investigate, mitigate, contain and remediate actual and suspected cybersecurity breaches and incidents could be significant. We may not be able to anticipate all types of security threats and implement preventive measures effective against all such threats. In addition, remote work has increased risks to our information technology systems and data, as our employees utilize network connections, computers and devices outside our premises or network, including working at home, while in transit and in public locations.
Cybersecurity threats and other disruptions, including the unauthorized, unlawful or accidental acquisition, modification, destruction, loss, alteration, encryption, disclosure of, or access to, our sensitive information or our information technology systems, or those of the third parties with whom we work, have become increasingly prevalent across industries, including the biotechnology industry, and may affect us in the future. From time to time, we have experienced cybersecurity events common to companies in our industry, such as phishing attempts, none of which has resulted in a material cybersecurity incident to date. However, there can be no assurance that we will not experience a material cybersecurity incident or other disruption in the future.
Additionally, despite our efforts to identify and address vulnerabilities in our information technology systems and infrastructure, we may not detect or remediate all vulnerabilities, including on a timely basis. We may also experience delays in developing and deploying remedial measures, security updates and patches designed to address identified vulnerabilities. Any such vulnerabilities could be exploited by threat actors and result in cybersecurity incidents or other disruptions.
The loss, corruption, unavailability of or damage to our data would interfere with and undermine the insights we draw from our City Platform and could impair the integrity of our clinical trial data, leading to potential regulatory delays or the inability to get our product candidates approved. If we do not accurately predict and identify our infrastructure requirements and failures and timely enhance our infrastructure, or if our remediation efforts are not successful, it could result in a material disruption of our business operations and development programs, including the loss or unauthorized disclosure of our sensitive information. A security breach or incident that leads to unauthorized acquisition, disclosure or other processing of our sensitive information could also affect our intellectual property rights and enable competitors to compete with us more effectively.
Likewise, as we work with third parties such as CROs, CDMOs, contractors and consultants, including for the manufacture of our product candidates and for the conduct of our clinical trials, similar events relating to their systems and operations could also have a material adverse effect on our business and lead to potential regulatory agency actions. Our ability to monitor these third parties’ information security practices is limited, and these third parties may not have adequate information security measures in place. For example, the loss of clinical trial data from completed, ongoing or future clinical trials could result in delays in or denials of our regulatory approval efforts and significantly increase our costs to recover or reproduce the data. Any security compromise affecting us, our collaborators or our industry, whether real or perceived, could harm our reputation, erode confidence in the effectiveness of our security measures, and lead to regulatory scrutiny. To the extent that any disruption or security breach were to result in a loss of, or damage to, our sensitive information or systems, or inappropriate disclosure of sensitive information, we could incur liability, our competitive position could be harmed, and the further development and commercialization of our product candidates could be delayed, result in
substantial costs and distract management. While we may be entitled to damages if the third parties with whom we work fail to satisfy their data privacy or security-related obligations to us, any award may be insufficient to cover our damages, or we may be unable to recover such award. In addition, supply-chain attacks have increased in frequency and severity, and we cannot guarantee that third parties’ infrastructure in our supply chain or that of the third parties with whom we work have not been compromised.
We may expend significant resources or modify our business activities (including our clinical trial activities) to try to protect against security incidents. Certain data privacy and security obligations have required us to implement and maintain specific security measures or industry-standard or reasonable security measures to protect our information technology systems and sensitive information.
Applicable data privacy and security obligations may require us, or we may voluntarily choose, to notify relevant stakeholders, including affected individuals, customers, regulators, and investors, of security incidents, or to take other actions. Such disclosures and related actions can be costly, and the disclosure or the failure to comply with such applicable requirements could lead to adverse consequences.
If we (or a third party with whom we work) experience failures, disruptions, security breaches and incidents, cyber-attacks we may experience material adverse consequences, such as government, enforcement measures, regulatory agency investigations, proceedings and other actions, penalties, fines, indemnification obligations, litigation (including class claims), negative publicity, monetary fund diversions, diversion of management attention, interruptions in our operations (including availability of data), financial loss, potential civil or criminal liability, collaborators’ loss of confidence, damage to our reputation and other consequences, which could materially adversely affect our business and results of operations.
Our contracts may not afford us limitations of liability, and even where they do, there can be no assurance that limitations of liability in our contracts are sufficient to protect us from liabilities, damages, or claims related to our data privacy and security obligations. Relatedly, our contracts with third parties with whom we work may limit the types and/or amounts of damages that we can recover from those third parties, even where the third party is responsible for a privacy or cybersecurity incident or violation. While we maintain insurance coverage for certain expenses and liabilities related to failures or breaches of our information technology systems, it may not be adequate or sufficient to cover all losses associated with such events. In addition, such insurance may not be available to us in the future on commercially reasonable terms or at all. Furthermore, if the information technology systems of third parties with whom we work become subject to disruptions or security breaches or incidents, we may have insufficient recourse against them.
In addition to experiencing a security incident, third parties may gather, collect, or infer sensitive information about us from public sources, data brokers, or other means that reveals competitively sensitive details about our organization and could be used to undermine our competitive advantage or market position. Additionally, sensitive information of ours, our customers, or clinical trial participants could be leaked, disclosed, or revealed as a result of or in connection with our employees’, personnel’s, or vendors’ use of generative AI technologies.
Issues relating to our current and planned use of AI in the identification of our programs and the engineering and development of our product candidates could adversely affect our business and operating results.
We currently incorporate certain AI solutions, among other technologies and capabilities, into our City Platform in a limited capacity, and we plan to expand our use of AI in our research and development activities in the future. There are risks involved in utilizing AI, including that AI-generated content, analyses, or recommendations we utilize could be deficient, that our competitors may more quickly or effectively adopt AI capabilities, or that our use of AI or other emerging technologies increases regulatory, cybersecurity and other significant risks. If any AI systems we use or develop fail to achieve their intended purposes — such as assisting in the identification of viable product candidates or targets, predicting biological outcomes, or producing reproducible results — our product development efforts
may be delayed or unsuccessful. If we are unable to successfully integrate and manage AI within our business as we expand our use of such technologies, or if AI fails to deliver the expected benefits, our ability to develop our programs and product candidates could be materially adversely affected.
Issues relating to the use of new and evolving technologies such as AI may cause us to experience brand or reputational harm, competitive harm, legal liability and new or enhanced governmental or regulatory scrutiny, and we may incur additional costs to resolve such issues. Known risks of AI generally include inaccuracy, hallucinations, bias, intellectual property infringement or misappropriation, data privacy and cybersecurity issues and data provenance disputes. Perceived or actual technical, legal, compliance, privacy, security, ethical or other issues relating to the use of AI in biopharmaceutical development may cause public confidence in AI to be undermined, which could slow market acceptance of product candidates discovered or developed with the assistance of AI. In addition, litigation or government regulation related to the use of AI may adversely impact our ability to identify programs and engineer and develop product candidates using AI, as well as increase the cost and complexity of doing so. For example, regulators may limit our ability to develop or implement AI models and algorithms and/or may eliminate or restrict the confidentiality of our proprietary technology, or may limit our ability to secure intellectual property rights to technologies created with the assistance of AI, which could have an adverse effect on our business, results of operations and financial condition.
We also face competition from other companies that use AI and related methods for drug engineering and development, some of which have more resources than we do and may have developed more effective methods than we and any third-party collaborators have, which may reduce our and any third-party collaborators’ effectiveness in identifying potential product candidates and attracting additional collaborators to work with us. In particular, biotechnology companies based in China present both known and emerging competitive threats to our business. Many of these companies operate within innovation ecosystems characterized by substantial government investment, access to large and rapidly expanding biological and clinical datasets, and accelerated regulatory or funding pathways. These factors may allow Chinese biotechnology companies to develop, train and deploy advanced computational models, drug discovery platforms or biologic design technologies more rapidly or at lower cost than we can. If our competitors are able to utilize new technologies more effectively (including but not limited to those that may involve AI or be created using AI) to discover, develop and commercialize products that compete with any of our programs and product candidates, such technologies could adversely impact our ability to compete.
Further, AI may have or produce errors or inadequacies that are not easily detectable. The quality of AI outputs depends heavily on the quality and quantity of input data. If the data used to train AI or the content, analyses or recommendations that AI applications assist in producing are or are alleged to be deficient, inaccurate, incomplete, hallucinatory, overbroad or biased, our business, financial condition and results of operations may be adversely affected. Developing, testing and deploying AI systems may also increase the cost profile of our operations due to the nature of the computing costs involved in such systems, which could adversely affect our business and operating results.
The legal landscape and legal protection for the use of AI remains uncertain, and the use of AI in drug discovery and development introduces evolving risks related to ownership, inventorship and protection of intellectual property generated by or with the assistance of AI technologies. For example, generative AI may be used improperly or inappropriately, which could lead to the tainting of our proprietary information and render us unable to qualify for certain patent or trade secret protection. Moreover, if our vendors, employees, suppliers or contractors with access to our proprietary and confidential information and know-how were to disclose such information as inputs to third-party AI tools, this could lead to loss of trade secret protection and otherwise impact our ability to realize the benefit of our intellectual property. If we do not have sufficient rights to collect or use the data on which any AI we utilize relies or to the outputs produced by our City Platform, we may incur liability through the alleged violation of certain laws, third-party privacy rights, online terms of service or other contracts to which we or our data providers are a party. In addition, we rely on third-party software and hardware for our City Platform. If the relevant software or hardware, or updates to such software or hardware, were to become unavailable to us in the future on reasonable commercial terms, or if they became the subject of allegations of intellectual property infringement, our ability to continue to use our City Platform could be
affected. Regulatory and legal frameworks governing inventions created with or using AI are still developing and may create uncertainty regarding our ability to secure and enforce rights in such inventions.
AI presents risks and challenges that can impact our business including regulatory, compliance, reputational, and competitive risks arising from our use of AI tools and evolving legal requirements.
Issues in the development and use of AI, combined with an uncertain regulatory environment, may result in reputational harm, liability or other adverse consequences to our business operations. As with many technological innovations, AI presents risks and challenges that could impact our business. As part of our City Platform, we use proprietary AI/ML molecular-design capabilities to support the discovery, design and engineering of RNAi trigger molecules and product candidates, and we may continue to expand our use of AI/ML and other computational tools in our research and development activities. Our vendors may also incorporate AI technologies into their products and services, including without specifically identifying the use of such technologies to us, and the providers of these AI technologies may not meet existing or evolving regulatory or industry standards with respect to data privacy, security or other legal requirements.
A growing number of federal, state, and international legislators, agencies and regulators are adopting laws and regulations and have focused enforcement efforts on the adoption of AI, and use of such technologies in compliance with ethical standards and societal expectations. These developments may increase our compliance burden and costs in connection with use of AI and lead to legal liability if we fail to meet evolving legal standards or if use of such technologies results in harms or other causes of action we did not predict. For example, the EU’s Artificial Intelligence Act (“AI Act”) may apply to us. This legislation imposes significant obligations on providers and deployers of AI systems and encourages providers and deployers of artificial intelligence systems to account for EU ethical principles in their development and use of these systems. In addition, various federal regulators have issued guidance and focused enforcement efforts on the use of AI in regulated sectors. The FDA, for example, issued draft guidance on the use of AI in regulatory decision-making for drug and biological products, which centers on the context of use while establishing a credibility assessment framework for establishing and evaluating AI model outputs intended to support regulatory decision-making. If we develop or use AI systems governed by these laws or regulations, including as informed by regulatory guidance, we would need to meet higher standards of data quality, transparency, monitoring and human oversight, and we would need to adhere to specific and potentially burdensome and costly ethical, accountability and administrative requirements, with the potential for significant enforcement or litigation in the event of any perceived non-compliance. We expect other jurisdictions will adopt similar laws. Uncertainty in the legal regulatory regime may require significant resources to modify and maintain business practices to comply with United States and non-United States laws, the nature of which cannot be determined at this time. Non-compliance or perceived non-compliance may lead to significant fines or significant restrictions on our ability to conduct our business activities.
We, and the third parties with whom we work are subject to a variety of stringent and evolving United States and foreign laws, regulations and rules, contractual obligations, industry standards, policies and other obligations related to data privacy and security. Any actual or perceived failure to comply with such obligations could expose us to regulatory investigations or actions, litigation (including class claims) and mass arbitration demands, significant fines or other penalties, disruptions of our business operations, reputational harm, loss of revenue or profits, loss of customers or sales, and other adverse business consequences.
In the ordinary course of our business, we and the third parties with whom we work process sensitive information. Our data processing activities subject us to numerous, evolving data privacy and security obligations, such as various laws, regulations, guidance, industry standards, external and internal privacy and security policies, contractual requirements and other obligations relating to data privacy and security.
The legislative and regulatory framework for the processing of sensitive information worldwide is rapidly evolving in a manner that is increasingly stringent and, globally, this legal and regulatory framework is likely to remain uncertain for the foreseeable future.
In the United States, numerous federal, state and local laws and regulations, including federal health information privacy laws, state information security and data breach notification laws, federal consumer protection laws (e.g., Section 5 of the Federal Trade Commission Act), state consumer protection and privacy laws and other similar laws (e.g., wiretapping and communications interception laws) govern the processing of health-related and other sensitive information. For example, HIPAA, as amended by the Health Information Technology for Economic and Clinical Health Act, imposes specific requirements relating to the privacy, security, and transmission of individually identifiable protected health information. At the state level, numerous U.S. states have enacted comprehensive privacy laws that impose certain obligations on covered businesses, including providing specific disclosures in privacy notices and affording individuals certain rights concerning their personal data. Similar laws are being considered in several other states, as well as at the federal and local levels, and we expect more states to pass similar laws in the future. While existing state comprehensive privacy laws exempt some data processed in the context of clinical trials, these developments may further complicate compliance efforts and increase legal risk and compliance costs for us and the third parties with whom we work.
Regulators and legislators in the United States are increasingly scrutinizing and restricting certain personal data transfers and transactions involving foreign countries. For example, the United States Department of Justice issued a rule entitled the Preventing Access to United States Sensitive Personal Data and Government-Related Data by Countries of Concern or Covered Persons, which places additional restriction on certain data transactions involving (e.g., China, Russia, Iran) and covered persons (e.g., individuals who are designated as such by the United States Attorney General or considered “foreign persona” and are majority owned by, organized under the laws of, a primary resident in, or a contractor of, a covered person or country of concern, as applicable) that may impact certain business activities such as vendor engagements, sale or sharing of data, employment of certain individuals, and investor agreements. Violations of the rule could lead to significant civil and criminal fines and penalties. The rule does not exempt key-coded or otherwise anonymized, pseudonymized, de-identified or encrypted data, which presents particular challenges for companies like ours and may impact our ability to engage in transactions or agreements with certain third parties in the future.
Outside the United States, an increasing number of laws, regulations and industry standards may govern data privacy and security. For example, the European Union’s General Data Protection Regulation (“EU GDPR”), the United Kingdom’s General Data Protection Regulation (“UK GDPR” and, together with the EU GDPR, “GDPR”) impose strict requirements for processing personal data including relating to processing of sensitive information (such as health data). Under GDPR, companies may face temporary or definitive bans on data processing and other corrective activities, fines of up to €20 million under the EU GDPR (£17.5 million GBP under the UK GDPR) or, in each case, 4% of annual global revenues, whichever is greater, and private litigation related to processing of personal data brought by classes of data subjects or consumer protection organizations authorized at law to represent their interests.
Our employees and personnel use generative AI and/or automated decision-making technologies to perform their work, and the disclosure and use of personal data in AI technologies is subject to various privacy laws and other privacy obligations. Governments have passed and are likely to pass additional laws and regulations regulating AI and/or automated decision-making technologies. Our use of this technology could result in additional compliance costs, regulatory investigations and actions, and lawsuits. If we are unable to use AI and/or automated decision-making technologies, it could make our business less efficient and result in competitive disadvantages.
In the ordinary course of business, we transfer personal data from Europe and other jurisdictions to the United States or other countries. However, we may be unable to transfer personal data from Europe and other jurisdictions to the United States or other countries due to data localization requirements or limitations on cross-border data flows. Europe and other jurisdictions have enacted laws requiring data to be localized or limiting the transfer of personal data to other countries. In particular, the European Economic Area (“EEA”) and the UK have significantly restricted the transfer of personal data to the United States and other countries. Other jurisdictions may adopt similarly stringent interpretations of their data localization and cross-border data transfer laws. Although there are currently various mechanisms
that may be used to transfer personal data from the EEA and UK to the United States in compliance with law, such as the EEA’s standard contractual clauses, the UK’s International Data Transfer Agreement / Addendum and the EU-United States Data Privacy Framework (“Framework”) and the UK extension thereto (which allows for transfers to relevant United States-based organizations who self-certify compliance and participate in the Framework), these mechanisms are subject to legal challenges, and there is no assurance that we can satisfy or rely on these measures to lawfully transfer personal data to the United States. If there is no lawful manner for us to transfer personal data from the EEA, the UK or other jurisdictions to the United States (or other countries), or if the requirements for a legally-compliant transfer are too onerous, we could face significant adverse consequences, including the interruption or degradation of our operations, the need to relocate part of or all of our business or data processing activities to other jurisdictions (such as Europe) at significant expense, increased exposure to regulatory actions, substantial fines and penalties, the inability to transfer data and work with partners, vendors and other third parties and injunctions against our processing or transferring of personal data necessary to operate our business. Additionally, companies that transfer personal data out of the EEA and UK to other jurisdictions, particularly to the United States, are subject to increased scrutiny from regulators, individual litigants and activities activist groups. Some European regulators have ordered certain companies to suspend or permanently cease certain transfers of personal data out of Europe for allegedly violating the GDPR’s cross-border data transfer limitations.
Additionally in the EEA, the NIS 2 Directive (“NIS 2”) regulates resilience and incident response capabilities of entities operating in a number of sectors, including the health sector. To the extent we are subject to NIS 2, we will require additional investment of our resources in compliance programs. Under NIS 2 companies may be subject to administrative fines of up to the higher amount of €10 million or 2% of worldwide turnover.
In addition to data privacy and security laws, we are contractually subject to industry standards adopted by industry groups and may become subject to such obligations in the future. We are also bound by other contractual obligations related to data privacy and security, and our efforts to comply with such obligations may not be successful. We publish privacy policies and other statements, such as compliance with certain certifications or self-regulatory principles, regarding data privacy and security. If these policies, materials or statements are found to be deficient, lacking in transparency, deceptive, unfair or misrepresentative of our practices, we may be subject to investigation, enforcement actions by regulators or other adverse consequences.
Obligations related to data privacy and security are quickly changing, becoming increasingly stringent, and creating uncertainty. Additionally, these obligations may be subject to differing applications and interpretations, which may be inconsistent or conflict among jurisdictions. Preparing for and complying with these obligations requires us to devote significant resources and may necessitate changes to our services, information technologies, systems and practices and to those of any third parties with whom we work.
We may at times fail (or be perceived to have failed) in our efforts to comply with our data privacy and security obligations. Moreover, despite our efforts, our personnel or third parties with whom we work, including CROs supporting our clinical trials, clinical trial sites with whom we have contracted and other third parties supporting our clinical trials, may fail to comply with such obligations, which could negatively impact our business operations. If we or the third parties with whom we work fail, or are perceived to have failed, to address or comply with applicable data privacy and security obligations, we could face significant consequences, including but not limited to: government enforcement actions (e.g., investigations, fines, penalties, audits, inspections and similar); litigation (including class-action claims), and mass arbitration demands; additional reporting requirements and/or oversight; bans on processing personal data; and orders to destroy or not use personal data. In particular, plaintiffs have become increasingly more active in bringing privacy-related claims against companies, including class claims and mass arbitration demands. Some of these claims allow for the recovery of statutory damages on a per violation basis, and, if viable, carry the potential for significant statutory damages, depending on the volume of data and the number of violations. Any of these events could have a material adverse effect on our reputation, business, financial condition, results of operations and prospects, including but not limited to: loss of customers; interruptions or stoppages in our business operations
(including, as relevant, clinical trials); inability to process personal data or to operate in certain jurisdictions; limited ability to develop or commercialize our products; expenditure of time and resources to defend any claim or inquiry; adverse publicity; or substantial changes to our business model or operations.
Our business could be affected by litigation, government investigations and enforcement actions.
We currently operate and plan to operate in a highly regulated industry and we could now or in the future be subject to litigation, government investigation and enforcement actions on a variety of matters in the United States or foreign jurisdictions, including, without limitation, intellectual property, regulatory, product liability, environmental, whistleblower, false claims, privacy, anti-kickback, anti-bribery, securities, commercial, employment and other allegations and legal proceedings which may arise from conducting our business. Any determination that our operations or activities are not in compliance with existing laws or regulations could result in the imposition of fines, civil and criminal penalties, equitable remedies, including disgorgement, injunctive relief and/or other sanctions against us, and remediation of any such findings could have an adverse effect on our business operations.
Legal proceedings, government investigations and enforcement actions can be expensive and time-consuming. An adverse outcome resulting from any such proceedings, investigations or enforcement actions could result in significant damages awards, fines, penalties, exclusion from the federal healthcare programs, healthcare debarment, injunctive relief, product recalls, reputational damage and modifications of our business practices, which could have a material adverse effect on our business and results of operations. Even if such a proceeding, investigation or enforcement action is ultimately decided in our favor, the investigation and defense thereof could require substantial financial and management resources and cause reputational harm.
Employee litigation and unfavorable publicity could negatively affect our future business.
Our employees may, from time to time, bring lawsuits against us regarding injury, creating a hostile workplace, discrimination, wage and hour disputes, sexual harassment or other employment issues. In recent years, there has been an increase in the number of discrimination and harassment allegations generally. Coupled with the expansion of social media platforms and similar devices that allow individuals access to a broad audience, these allegations have had a significant negative impact on some businesses. Certain companies that have faced employment- or harassment-related lawsuits have had to terminate management or other key personnel and have suffered reputational harm that has negatively impacted their business. If we were to face any employment-related allegations, our business could be negatively affected.
Unfavorable United States or global economic conditions could adversely affect our business, financial condition or results of operations.
Our results of operations could be adversely affected by general conditions in the global economy and financial markets. The global economy and financial markets have experienced extreme volatility and disruptions, including severely diminished liquidity and credit availability, declines in consumer confidence, rising inflation, uncertainty from changes in tariff policies, fluctuating interest rates, declines in economic growth, global supply chain disruptions and uncertainty about economic stability. The global economy and financial markets may also be adversely affected by the current or anticipated impact of military conflict, terrorism or other geopolitical events, including the ongoing war in Ukraine, the Israel-Gaza conflict and the increasingly strained relationship between the United States and China. Sanctions imposed by the United States and other countries in response to such conflicts may adversely impact the financial markets and the global economy, and the economic countermeasures by the affected countries or others could exacerbate market and economic instability.
There can be no assurance that further deterioration in credit and financial markets and confidence in economic conditions will not occur. A severe or prolonged economic downturn could result in a variety of risks to our business, including weakened demand for any product candidates or products we may develop and our ability to raise additional capital when needed on acceptable terms, if at all. A weak or
declining economy could also strain our suppliers, possibly resulting in supply disruption. If the equity and credit markets deteriorate, it may make any necessary equity or debt financing more difficult, more costly and more dilutive. Failure to secure any necessary financing in a timely manner and on favorable terms could impair our ability to achieve our growth strategy, could harm our financial performance and stock price and could require us to delay or abandon clinical development plans. In addition, there is a risk that our current or future service providers, CDMOs or other collaborators may not survive such difficult economic times, which could directly affect our ability to attain our operating goals on schedule and on budget. We cannot anticipate all of the ways in which the current economic climate and financial market conditions could adversely impact our business.
A decline in the federal budget, changes in spending or budgetary priorities of the United States government, a prolonged United States government shutdown or delays in contract awards may significantly and adversely affect our future revenues, cash flow and financial results.
In recent years, United States government appropriations have been affected by larger United States government budgetary issues and related legislation. As a result, the Department of Defense funding levels have fluctuated and have been difficult to predict. Future spending levels are subject to a wide range of factors, including Congressional action. In addition, in the past, United States debt ceiling and budget deficit concerns have increased the possibility of additional credit-rating downgrades and economic slowdowns, or a recession in the United States. Although United States lawmakers passed legislation to raise the federal debt ceiling on multiple occasions, ratings agencies have lowered or threatened to lower the long-term sovereign credit rating on the United States. The impact of this or any further downgrades to the United States government’s sovereign credit rating or its perceived creditworthiness could adversely affect the United States and global financial markets and economic conditions.
As a result, government spending levels are difficult to predict beyond the near term due to numerous factors, including the external threat environment, future government priorities and the state of government finances. Significant changes in government spending or changes in United States government priorities, policies and requirements could have a material adverse effect on our results of operations, financial condition or liquidity.
Adverse developments affecting the financial services industry, such as actual events or concerns involving liquidity, defaults, or non-performance by financial institutions or transactional counterparties, could adversely affect our current and projected business operations and our financial condition and results of operations.
Actual events involving limited liquidity, defaults, non-performance or other adverse developments that affect financial institutions, transactional counterparties or other companies in the financial services industry or the financial services industry generally, or concerns or rumors about any events of these kinds or other similar risks, have in the past and may in the future lead to market-wide liquidity problems.
Although we assess our banking and customer relationships as we believe necessary or appropriate, our access to funding sources and other credit arrangements in amounts adequate to finance or capitalize our current and projected future business operations could be significantly impaired by factors that affect our Company, the financial institutions with which we have credit agreements or arrangements directly, or the financial services industry or economy in general. These factors could include, among others, events such as liquidity constraints or failures, the ability to perform obligations under various types of financial, credit or liquidity agreements or arrangements, disruptions or instability in the financial services industry or financial markets, or concerns or negative expectations about the prospects for companies in the financial services industry.
The results of events or concerns that involve one or more of these factors could include a variety of material and adverse impacts on our current and projected business operations and our financial condition and results of operations. These could include, but may not be limited to, the following:
•
delayed access to deposits or other financial assets or the uninsured loss of deposits or other financial assets;
•
potential or actual breach of statutory, regulatory or contractual obligations, including obligations that require us to maintain letters of credit or other credit support arrangements; and
•
termination of cash management arrangements and/or delays in accessing or actual loss of funds subject to cash management arrangements.
In addition, investor concerns regarding the United States or international financial systems could result in less favorable commercial financing terms, including higher interest rates or costs and tighter financial and operating covenants, or systemic limitations on access to credit and liquidity sources, thereby making it more difficult for us to acquire financing on acceptable terms or at all. Any decline in available funding or access to our cash and liquidity resources could, among other risks, adversely impact our ability to meet our operating expenses, financial obligations or fulfill our other obligations, result in breaches of our financial and/or contractual obligations or result in violations of federal or state wage and hour laws. Any of these impacts, or any other impacts resulting from the factors described above or other related or similar factors not described above, could have material adverse impacts on our liquidity, our current and/or planned business operations, and our current or projected financial condition and results of operations.
In addition, any further deterioration in the macroeconomic economy or financial services industry could lead to losses or defaults by our suppliers, which in turn, could have a material adverse effect on our current and/or planned business operations and our current or projected results of operations and financial condition. For example, a customer may fail to make payments when due, default under their agreements with us, become insolvent or declare bankruptcy, or a supplier may determine that it will no longer deal with us as a customer. In addition, a customer or supplier could be adversely affected by any of the liquidity or other risks that are described above as factors that could result in material adverse impacts on the Company, including but not limited to delayed access or loss of access to uninsured deposits or loss of the ability to draw on existing credit facilities involving a troubled or failed financial institution. Any customer, collaborator or supplier bankruptcy or insolvency, or the failure of any customer or collaborator to make payments when due, or any breach or default by a customer, collaborator or supplier, or the loss of any significant supplier or collaborator relationships, could result in material losses to the Company and may have a material adverse impact on our business.
The increasing use of social media platforms presents risks and challenges.
We and our employees are increasingly utilizing social media tools as a means of communication both internally and externally. Despite our efforts to monitor evolving social media communication guidelines and comply with applicable rules, there is risk that the use of social media by us or our employees to communicate about our product candidates, operations, or business may cause us to be found in violation of applicable legal or contractual requirements. In addition, our employees may knowingly or inadvertently make use of social media in ways that may not comply with our social media policy or other legal or contractual requirements, which may give rise to liability, lead to the loss of trade secrets or other intellectual property, or result in public exposure of personal information of our employees, clinical trial patients, collaboration partners, and others, and which could have an adverse effect on our business, financial conditions and results of operations. Furthermore, negative posts or comments about us or our product candidates in social media could seriously damage our reputation, brand image and goodwill.
In addition, it is possible for individuals or groups to target companies with disruptive social media campaigns related to a request for access to unapproved drugs for patients with significant unmet medical need. If we experience a similar social media campaign regarding our decision to provide or not provide access to our product candidates under an expanded access policy, our reputation may be negatively affected and our business may be harmed.
CAUTIONARY NOTE REGARDING FORWARD-LOOKING STATEMENTS
This prospectus, including the sections entitled “Prospectus Summary,” “Risk Factors,” “Management’s Discussion and Analysis of Financial Condition and Results of Operations,” and “Business,” contains express or implied forward-looking statements that are based on our management’s belief and assumptions and on information currently available to our management. Although we believe that the expectations reflected in these forward-looking statements are reasonable, these statements relate to future events or our future operational or financial performance, and involve known and unknown risks, uncertainties, and other factors that may cause our actual results, performance or achievements to be materially different from any future results, performance or achievements expressed or implied by these forward-looking statements. Forward-looking statements in this prospectus include, but are not limited to, statements about:
•
advance our product candidates through preclinical and clinical development, including continuing the Phase 1 clinical trial for CITY-FXI and advancing other programs through preclinical and clinical development;
•
the initiation, timing, progress and results of our research and development programs, preclinical studies and clinical trials for our current or future product candidates and effectiveness of our City Platform;
•
the ability of clinical trials to demonstrate safety and efficacy of our product candidates, and other positive results, and the ability of our preclinical studies and earlier clinical trials to predict later clinical trial results;
•
the timing, scope and likelihood of regulatory filings and approvals of our product candidates;
•
the implementation of our business model, and strategic plans for our business, programs, and current and future product candidates;
•
our ability to obtain additional cash and the sufficiency of our existing cash and cash equivalents to fund our future operating expenses and capital expenditure requirements;
•
the accuracy of our estimates regarding expenses, future revenue, capital requirements and needs for additional financing;
•
the size and growth potential of the markets for our product candidates, and our ability to serve those markets;
•
our potential and ability to successfully manufacture and supply our current and future product candidates for clinical trials and for commercial use, if approved;
•
our ability to leverage programs within our initial target indications and to progress additional programs to further develop our pipeline;
•
the scope of protection we are able to establish and maintain for intellectual property rights covering our product candidates;
•
developments relating to our competitors and our industry, including competing product candidates and RNAi therapeutics;
•
developments related to our proprietary City Platform;
•
existing regulations and regulatory developments in the United States and other jurisdictions;
•
our ability to obtain funding for our operations necessary to complete further development and commercialization of our product candidates;
•
expectations regarding future events under collaboration and licensing agreements, including potential future payments, as well as our plans and strategies for entering into further collaboration and licensing agreements;
•
general economic, industry and market conditions, including fluctuating interest rates and inflationary pressures;
•
our ability to attract and retain key scientific and management personnel;
•
our expectations regarding the period during which we will qualify as an emerging growth company under the JOBS Act; and
•
our anticipated use of our existing cash and cash equivalents and the proceeds from this offering.
In some cases, forward-looking statements can be identified by terminology such as “may,” “should,” “expects,” “intends,” “plans,” “anticipates,” “believes,” “estimates,” “predicts,” “potential,” “continue,” or the negative of these terms or other comparable terminology. These statements are only predictions. You should not place undue reliance on forward-looking statements because they involve known and unknown risks, uncertainties and other factors, which are, in some cases, beyond our control and which could materially affect results. Factors that may cause actual results to differ materially from current expectations include, among other things, those listed under the section entitled “Risk Factors” and elsewhere in this prospectus. If one or more of these risks or uncertainties occur, or if our underlying assumptions prove to be incorrect, actual events or results may vary significantly from those implied or projected by the forward-looking statements. No forward-looking statement is a guarantee of future performance. You should read this prospectus and the documents that we reference in this prospectus and have filed with the SEC as exhibits to the registration statement, of which this prospectus is a part, completely and with the understanding that our actual future results may be materially different from any future results expressed or implied by these forward-looking statements.
The forward-looking statements in this prospectus represent our views as of the date of this prospectus. We anticipate that subsequent events and developments will cause our views to change. In addition, statements that “we believe” and similar statements reflect our beliefs and opinions on the relevant subject. These statements are based upon information available to us as of the date of this prospectus, and while we believe such information forms a reasonable basis for such statements, such information may be limited or incomplete, and our statements should not be read to indicate that we have conducted an exhaustive inquiry into, or review of, all potentially available relevant information. These statements are inherently uncertain, and you are cautioned not to unduly rely upon these statements.
Except as required by applicable law, we do not plan to publicly update or revise any forward-looking statements contained herein until after we distribute this prospectus, whether as a result of any new information, future events or otherwise. You should therefore not rely on these forward-looking statements as representing our views as of any date subsequent to the date of this prospectus.
This prospectus includes statistical and other industry and market data that we obtained from industry publications and research, surveys and studies conducted by third parties. Industry publications and third-party research, surveys and studies generally indicate that their information has been obtained from sources believed to be reliable, although they do not guarantee the accuracy or completeness of such information. We are responsible for all of the disclosure contained in this prospectus, and we believe that these sources are reliable; however, we have not independently verified the information contained in such publications.
USE OF PROCEEDS
We estimate that our net proceeds from the sale of 9,722,222 shares of our common stock in this offering will be approximately $158.3 million (or approximately $182.7 million if the underwriters exercise their option to purchase additional shares of our common stock in full), assuming an initial public offering price of $18.00 per share, the midpoint of the estimated offering price range set forth on the cover page of this prospectus, and after deducting underwriting discounts and commissions and estimated offering expenses payable by us.
A $1.00 increase (decrease) in the assumed initial public offering price of $18.00 per share, the midpoint of the estimated offering price range set forth on the cover page of this prospectus, would increase (decrease) our net proceeds from this offering by $9.0 million, assuming the number of shares offered by us, as set forth on the cover page of this prospectus, remains the same and after deducting underwriting discounts and commissions and estimated offering expenses payable by us. A 1.0 million share increase (decrease) in the number of shares offered by us, as set forth on the cover page of this prospectus, would increase (decrease) our net proceeds from this offering by $16.7 million, assuming no change in the assumed initial public offering price per share, the midpoint of the estimated offering price range set forth on the cover page of this prospectus, and after deducting underwriting discounts and commissions and estimated offering expenses payable by us. The information discussed above is illustrative only and will adjust based on the actual initial public offering price and other terms of this offering determined at pricing.
The principal purposes of this offering are to create a public market for our common stock and thereby facilitate future access to the public equity markets, increase our visibility in the marketplace and obtain additional capital. We currently intend to use the net proceeds from this offering, together with our existing cash and cash equivalents, as follows:
•
approximately $30.0 million to fund the clinical development of CITY-FXI, including our ongoing Phase 1 clinical trial and the initiation of a Phase 2 TKA study;
•
approximately $20.0 million to fund the clinical development of CITY-RBP4, including our Phase 1 clinical trial and preparations for later-stage clinical development for the treatment of Stargardt disease and GA;
•
approximately $40.0 million to fund the clinical development of CITY-TFR2, including an IIT in China and the Phase 1/2 clinical trial, both focused on the treatment of anemia associated with myelofibrosis;
•
approximately $70.0 million to advance our City Platform, including continued investment to advance cityRNA triggers (CITY-O02 and CITY-C02) to the pipeline, as well as additional preclinical product candidates toward IND or IND-equivalent submission and clinical development; and
•
the remainder for working capital and other general corporate purposes.
We believe, based on our current operating plan, that the net proceeds from this offering, together with our existing cash and cash equivalents, will be sufficient to fund our operations into late 2028. We have based this estimate on our current assumptions, which may prove to be wrong, and we may exhaust our available capital resources sooner than we expect.
Our expected use of the net proceeds from this offering represents our intentions based on our current plans and business conditions. As of the date of this prospectus, we cannot predict with certainty all of the particular uses for the net proceeds to be received upon the completion of this offering or the amounts that we will actually spend on the uses set forth above. We expect that we will require additional capital in order to fund the development of our product candidates beyond the specific activities described above, and we will require substantial additional capital to complete later stages of clinical development of, obtain regulatory approval for, and commercialize any of our product candidates. We may also use a portion of the net proceeds to in-license, acquire or invest in complementary businesses or technologies to continue to build our pipeline, research and development capabilities and our
intellectual property position, although we currently have no agreements, commitments or understandings with respect to any such transaction.
Due to the many inherent uncertainties in the development of our programs and product candidates, the amounts and timing of our actual expenditures may vary significantly depending on numerous factors, including the progress of our research and development, timing of patient enrollment and evolving regulatory requirements, the timing and success of preclinical studies, our ongoing or future clinical trials, the timing of regulatory submissions, any strategic alliances that we may enter into with third parties or strategic opportunities that become available to us, and other factors described in “Risk Factors” in this prospectus, as well as the amount of cash used in our operations and any unforeseen cash needs.
Our management will retain broad discretion in the application of the net proceeds we receive from our initial public offering, and investors will be relying on the judgment of our management regarding the application of the net proceeds. Pending our use of the net proceeds from this offering, we intend to invest the net proceeds in a variety of capital preservation instruments, including short-term and long-term interest-bearing instruments, investment-grade securities, and direct or guaranteed obligations of the United States government. We cannot predict whether the proceeds invested will yield a favorable return.
DIVIDEND POLICY
We have never declared or paid cash dividends on our capital stock. We currently intend to retain all available funds and any future earnings to fund the growth and development of our business. We do not intend to declare or pay cash dividends to our stockholders in the foreseeable future. Any future determination to declare dividends will be made at the discretion of our board of directors and will depend on our financial condition, operating results, capital requirements, general business conditions, and other factors that our board of directors may deem relevant.
In addition, our ability to pay cash dividends on our capital stock in the future may be limited by the terms of any future debt or preferred securities we issue or any credit facilities we enter into.
CAPITALIZATION
The following table sets forth our cash, cash equivalents, and restricted cash and total capitalization as of June 30, 2026:
•
on an actual basis;
•
on a pro forma basis, giving effect to (i) the automatic conversion of all outstanding shares of convertible preferred stock into an aggregate of 28,848,641 shares of common stock immediately prior to the completion of this offering, (ii) the automatic conversion of the outstanding principal and accrued interest under the Biogen Note into an aggregate of 2,133,788 shares of our common stock immediately prior to the completion of this offering, at a conversion price equal to 85% of the assumed initial public offering price of $18.00 per share, the midpoint of the estimated offering price range listed on the cover page of this prospectus, including a change in the fair value of the Biogen Note liability of $5.3 million based on the assumed initial public offering price of $18.00 per share, and (iii) the filing and effectiveness of our amended and restated certificate of incorporation, which will occur immediately prior to the completion of this offering; and
•
on a pro forma as adjusted basis, giving effect to (i) the pro forma adjustments set forth above and (ii) the issuance and sale of 9,722,222 shares of common stock in this offering at the assumed initial public offering price of $18.00 per share, the midpoint of the estimated offering price range listed on the cover page of this prospectus, after deducting underwriting discounts and commissions and estimated offering expenses payable by us.
The pro forma as adjusted information below is illustrative only, and our capitalization following the completion of this offering will be adjusted based on the actual initial public offering price and other terms of this offering determined at pricing.
The following table should be read together with “Management’s Discussion and Analysis of Financial Condition and Results of Operations,” “Description of Capital Stock,” and the consolidated financial statements and related notes appearing elsewhere in this prospectus.
| |
|
|
As of June 30, 2026
|
|
|
(in thousands, except share and per share data)
|
|
|
Actual
|
|
|
Pro Forma
|
|
|
Pro Forma As
Adjusted(1)
|
|
|
Cash, cash equivalents, and restricted cash
|
|
|
|
$ |
172,109 |
|
|
|
|
$ |
172,109 |
|
|
|
|
$ |
330,583 |
|
|
|
Convertible notes, net (including current portion)
|
|
|
|
$ |
33,089 |
|
|
|
|
$ |
— |
|
|
|
|
$ |
— |
|
|
|
Convertible preferred stock (Series Seed, A, B, and B-1), par value $0.0001 per share; 26,266,656 shares authorized, issued, and outstanding, actual; no shares authorized, issued, and outstanding, pro forma and pro forma as adjusted
|
|
|
|
|
256,056 |
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
Stockholders’ deficit:
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
Preferred stock, par value $0.0001 per share; no shares
authorized, issued, and outstanding, actual; 10,000,000
shares authorized, no shares issued or outstanding,
pro forma and pro forma as adjusted
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
Common stock, par value $0.0001 per share; 42,350,000
shares authorized, 9,627,628 shares issued, 7,663,207
shares outstanding, actual; 500,000,000 shares
authorized, 40,610,057 shares issued, 38,645,636
shares outstanding, pro forma; 500,000,000 shares
authorized, 50,332,279 shares issued, 48,367,858
shares outstanding, pro forma as adjusted
|
|
|
|
|
— |
|
|
|
|
|
3 |
|
|
|
|
|
4 |
|
|
|
Additional paid-in capital
|
|
|
|
|
10,830 |
|
|
|
|
|
305,291 |
|
|
|
|
|
463,540 |
|
|
|
Accumulated deficit
|
|
|
|
|
(142,547) |
|
|
|
|
|
(147,866) |
|
|
|
|
|
(147,866) |
|
|
|
Total stockholders’ (deficit) equity
|
|
|
|
$ |
(131,717) |
|
|
|
|
|
157,428 |
|
|
|
|
|
315,678 |
|
|
|
Total capitalization
|
|
|
|
$ |
157,428 |
|
|
|
|
$ |
157,428 |
|
|
|
|
$ |
315,678 |
|
|
(1)
Each $1.00 increase or decrease, as applicable, in the assumed initial public offering price of $18.00 per share, the midpoint of the estimated offering price range listed on the cover page of this prospectus for this offering, would increase or decrease, as applicable, the pro forma as adjusted amount of each of cash, cash equivalents, and restricted cash, additional paid-in capital, total stockholders’ deficit and total capitalization by $9.0 million, assuming that the number of shares offered by us in this offering remains the same and after deducting underwriting discounts and commissions and estimated offering expenses payable by us. Similarly, each increase or decrease, as applicable, of 1.0 million shares in the number of shares of common stock offered by us at the assumed initial public offering price per share, the midpoint of the estimated offering price range listed on the cover page of this prospectus in this offering, would increase or decrease, as applicable, the pro forma as adjusted amount of each of cash, cash equivalents, and restricted cash, additional paid-in capital, total stockholders’ deficit and total capitalization by $16.7 million, after deducting underwriting discounts and commissions and estimated offering expenses payable by us.
The number of shares of common stock that will be outstanding after this offering on a pro forma and pro forma as adjusted basis is based on 40,610,057 shares of common stock (which includes 1,964,421 shares of restricted common stock subject to repurchase or forfeiture) outstanding as of June 30, 2026 after giving effect to the automatic conversion of all outstanding shares of our convertible preferred stock and the Biogen Note into an aggregate of 30,982,429 shares of our common stock immediately prior to the completion of this offering, and excludes:
•
5,404,513 shares of common stock issuable upon exercise of outstanding stock options as of June 30, 2026 under our 2023 Plan, with a weighted-average exercise price of $1.97 per share;
•
988,458 shares of common stock issuable upon exercise of outstanding stock options granted after June 30, 2026 under our 2023 Plan, with a weighted-average exercise price of $8.64 per share;
•
1,482,874 shares of common stock reserved for future issuance as of June 30, 2026 under our 2023 Plan, which will cease to be available for issuance at the time that our 2026 Plan becomes effective;
•
6,516,101 shares of our common stock reserved for future issuance under our 2026 Plan, which will become effective on the date immediately prior to the effectiveness of the registration statement of which this prospectus forms a part, as well as any automatic increases in the number of shares of common stock reserved for future issuance under the 2026 Plan and any shares underlying outstanding stock awards granted under our 2023 Plan that expire or are repurchased, forfeited, cancelled or withheld; and
•
503,322 shares of common stock reserved for future issuance under our ESPP, which will become effective on the date immediately prior to the effectiveness of the registration statement of which this prospectus forms a part, as well as any automatic increases in the number of shares of common stock reserved for future issuance under the ESPP.
DILUTION
If you invest in our common stock in this offering, your ownership interest will be diluted to the extent of the difference between the initial public offering price per share of our common stock and the pro forma as adjusted net tangible book value per share of our common stock immediately after this offering.
Our historical net tangible book deficit as of June 30, 2026 was a deficit of $132.6 million, or $17.31 per share of our common stock. Our historical net tangible book deficit represents the amount of our total tangible assets (net of deferred offering costs) less our total liabilities and convertible preferred stock. Historical net tangible book deficit per share represents historical net tangible book deficit divided by the number of shares of our common stock outstanding as of June 30, 2026 (which excludes 1,964,421 shares of restricted common stock subject to repurchase or forfeiture).
Our pro forma net tangible book value as of June 30, 2026 was $156.5 million, or $4.05 per share. Pro forma net tangible book value per share represents the amount of our total tangible assets (net of deferred offering costs) less our total liabilities, divided by the number of shares of our common stock outstanding as of June 30, 2026, after giving effect to (i) the automatic conversion of all outstanding shares of convertible preferred stock into an aggregate of 28,848,641 shares of our common stock immediately prior to the completion of this offering and (ii) the automatic conversion of the outstanding principal and accrued interest under the Biogen Note into an aggregate of 2,133,788 shares of our common stock immediately prior to the completion of this offering, at a conversion price equal to 85% of the assumed initial public offering price of $18.00 per share, the midpoint of the estimated offering price range listed on the cover page of this prospectus.
After giving further effect to the sale of 9,722,222 shares of common stock that we are offering at the assumed initial public offering price of $18.00 per share, the midpoint of the estimated offering range set forth on the cover page of this prospectus, and after deducting underwriting discounts and commissions and estimated offering expenses payable by us, our pro forma as adjusted net tangible book value as of June 30, 2026 would have been $315.7 million, or $6.53 per share. This amount represents an immediate increase in pro forma as adjusted net tangible book value of $2.48 per share to our existing stockholders and an immediate dilution in pro forma as adjusted net tangible book value of $11.47 per share to new investors purchasing shares of common stock in this offering.
Dilution per share to new investors is determined by subtracting pro forma as adjusted net tangible book value per share after this offering from the initial public offering price per share paid by new investors. The following table illustrates this dilution (without giving effect to any exercise by the underwriters of their option to purchase additional shares):
| |
Assumed initial public offering price per share
|
|
|
|
|
|
|
|
|
|
$ |
18.00 |
|
|
| |
Historical net tangible book deficit per share as of June 30, 2026
|
|
|
|
$ |
(17.31) |
|
|
|
|
|
|
|
|
| |
Increase in net tangible book value per share attributable to the pro forma adjustment described above
|
|
|
|
|
21.36 |
|
|
|
|
|
|
|
|
| |
Pro forma net tangible book value per share as of June 30, 2026
|
|
|
|
|
4.05 |
|
|
|
|
|
|
|
|
| |
Increase in pro forma net tangible book value per share attributable to new investors participating in this offering
|
|
|
|
|
2.48 |
|
|
|
|
|
|
|
|
| |
Pro forma as adjusted net tangible book value per share after this offering
|
|
|
|
|
|
|
|
|
|
|
6.53 |
|
|
| |
Dilution per share to new investors participating in this offering
|
|
|
|
|
|
|
|
|
|
$ |
11.47 |
|
|
The dilution information discussed above is illustrative only and may change based on the actual initial public offering price and other terms of this offering. Each $1.00 increase or decrease, as applicable, in the assumed initial public offering price of $18.00 per share, which is the midpoint of the estimated offering price range set forth on the cover page of this prospectus, would increase or decrease, as applicable, the pro forma as adjusted net tangible book value per share after this offering by $0.18, and dilution in pro forma net tangible book value per share to new investors by $0.82, assuming that the number of shares of common stock offered by us, as set forth on the cover page of this prospectus,
remains the same, and after deducting underwriting discounts and commissions and estimated offering expenses payable by us. Similarly, each increase or decrease, as applicable, of 1.0 million shares in the number of shares of common stock offered by us, as set forth on the cover page of this prospectus, would increase or decrease, as applicable, our pro forma as adjusted net tangible book value per share after this offering by $0.20 per share, in each case, and decrease or increase, as applicable, the dilution to investors participating in this offering by $0.20 per share, assuming that the assumed initial public offering price, the midpoint of the estimated offering price range set forth on the cover page of this prospectus, remains the same, and after deducting underwriting discounts and commissions and estimated offering expenses payable by us.
If the underwriters exercise their option to purchase up to 1,458,333 additional shares of our common stock in full, the pro forma as adjusted net tangible book value after the offering would be $6.83 per share, the increase in pro forma as adjusted net tangible book value per share to existing stockholders would be $2.78 per share and the dilution per share to new investors would be $11.17 per share, in each case assuming an initial public offering price of $18.00 per share, the midpoint of the estimated offering range set forth on the cover page of this prospectus.
The following table summarizes on the pro forma as adjusted basis described above, as of June 30, 2026, the differences between the number of shares of common stock purchased from us by our existing stockholders and common stock by new investors purchasing shares in this offering, the total consideration paid to us in cash and the average price per share paid by existing stockholders for shares of common stock issued prior to this offering and the price to be paid by new investors for shares of common stock in this offering. The calculation below is based on the assumed initial public offering price of $18.00 per share, the midpoint of the estimated offering price range set forth on the cover page of this prospectus, before deducting underwriting discounts and commissions and estimated offering expenses payable by us.
| |
|
|
Shares Purchased
|
|
|
Total Consideration
|
|
|
Weighted-
Average
Price Per
Share
|
|
| |
|
|
Number
|
|
|
Percentage
|
|
|
Amount
|
|
|
Percentage
|
|
| |
|
|
(in thousands, except share, per share and percent data)
|
|
|
Existing stockholders before this
offering
|
|
|
|
|
38,645,636 |
|
|
|
|
|
79.9% |
|
|
|
|
$ |
286,801 |
|
|
|
|
|
62.1% |
|
|
|
|
$ |
7.42 |
|
|
|
New investors purchasing shares in this offering
|
|
|
|
|
9,722,222 |
|
|
|
|
|
20.1% |
|
|
|
|
|
175,000 |
|
|
|
|
|
37.9% |
|
|
|
|
$ |
18.00 |
|
|
|
Total
|
|
|
|
|
48,367,858 |
|
|
|
|
|
100.0% |
|
|
|
|
$ |
461,801 |
|
|
|
|
|
100.0% |
|
|
|
|
|
|
|
|
The table above assumes no exercise of the underwriters’ option to purchase additional shares in this offering. If the underwriters’ option to purchase additional shares is exercised in full, the number of shares of our common stock held by existing stockholders would be reduced to 77.6% of the total number of shares of our common stock outstanding after this offering, and the number of shares of our common stock held by new investors purchasing common stock in this offering would be increased to 22.4% of the total number of shares of our common stock outstanding after this offering.
Each $1.00 increase or decrease, as applicable, in the assumed initial public offering price of $18.00 per share, which is the midpoint of the estimated offering price range set forth on the cover page of this prospectus, would increase or decrease, as applicable, the total consideration paid by new investors by $9.7 million and, in the case of an increase, would increase the percentage of total consideration paid by new investors to 39.2% and, in the case of a decrease, would decrease the percentage of total consideration paid by new investors to 36.6%, assuming that the number of shares offered by us, as set forth on the cover page of this prospectus, remains the same. Similarly, each increase or decrease, as applicable, of 1.0 million shares in the number of shares offered by us, would increase or decrease, as applicable, the total consideration paid by new investors by $18.0 million and, in the case of an increase, would increase the percentage of total consideration paid by new investors to 40.2% and, in the case of a decrease, would decrease the percentage of total consideration
paid by new investors to 35.4%, assuming that the assumed initial public offering price of $18.00 per share, the midpoint of the estimated offering price range set forth on the cover page of this prospectus, remains the same.
The foregoing tables and calculations (other than the historical net tangible book deficit calculation) are based on 38,645,636 shares of common stock (which excludes 1,964,421 shares of restricted common stock subject to repurchase or forfeiture) outstanding as of June 30, 2026, after giving effect to the automatic conversion of outstanding shares of our convertible preferred stock and the Biogen Note into shares of common stock immediately prior to the completion of this offering, and excludes:
•
5,404,513 shares of common stock issuable upon exercise of outstanding stock options as of June 30, 2026 under our 2023 Plan, with a weighted-average exercise price of $1.97 per share;
•
988,458 shares of common stock issuable upon exercise of outstanding stock options granted after June 30, 2026 under our 2023 Plan, with a weighted-average exercise price of $8.64 per share;
•
1,482,874 shares of common stock reserved for future issuance as of June 30, 2026 under our 2023 Plan, which will cease to be available for issuance at the time that our 2026 Plan becomes effective;
•
6,516,101 shares of our common stock reserved for future issuance under our 2026 Plan, which will become effective on the date immediately prior to the effectiveness of the registration statement of which this prospectus forms a part, as well as any automatic increases in the number of shares of common stock reserved for future issuance under the 2026 Plan and any shares underlying outstanding stock awards granted under our 2023 Plan that expire or are repurchased, forfeited, cancelled or withheld; and
•
503,322 shares of common stock reserved for future issuance under our ESPP, which will become effective on the date immediately prior to the effectiveness of the registration statement of which this prospectus forms a part, as well as any automatic increases in the number of shares of common stock reserved for future issuance under the ESPP.
To the extent that outstanding options are exercised, or we issue additional shares of common stock in the future, there will be further dilution to new investors. In addition, we may choose to raise additional capital because of market conditions or strategic considerations, even if we believe that we have sufficient funds for our current or future operating plans. If we raise additional capital through the sale of equity or convertible debt securities, the issuance of these securities could result in further dilution to our stockholders.
MANAGEMENT’S DISCUSSION AND ANALYSIS OF FINANCIAL CONDITION AND
RESULTS OF OPERATIONS
You should read the following discussion and analysis of our financial condition and results of operations in conjunction with our audited consolidated financial statements and unaudited condensed consolidated financial statements and related notes, as well as other financial information included elsewhere in this prospectus. This discussion and analysis and other parts of this prospectus contain forward-looking statements based upon our current plans and expectations that involve risks, uncertainties and assumptions, such as statements regarding our plans, strategies, objectives, expectations, intentions, and beliefs. Our actual results and the timing of events could differ materially from those anticipated in these forward-looking statements as a result of various factors, including those set forth under “Risk Factors” and elsewhere in this prospectus. You should carefully read the “Risk Factors” section of this prospectus to gain an understanding of the important factors that could cause actual results to differ materially from our forward-looking statements. Our historical results are not necessarily indicative of the results that may be expected for any period in the future. Please also see the section titled “Cautionary Note Regarding Forward-Looking Statements.” Unless otherwise indicated, all references in this section to the “company,” “we,” “our,” “us,” or similar terms refer to City Therapeutics, Inc. and its wholly owned subsidiary.
Overview
We are a clinical-stage biotechnology company developing the next-generation of RNAi therapeutics designed to silence the expression of disease-relevant proteins, with the aim of delivering meaningful and durable benefits for patients in need. The City Platform is the engine behind every therapeutic we develop. Our City Platform is designed to integrate three core capabilities: RNAi trigger engineering, novel delivery conjugates designed to enable tissue selectivity, and target discovery and selection enabled by our insights into human genetics. Critical to every RNAi medicine is the trigger that harnesses the cellular RNAi machinery to silence a disease-relevant gene. Our most advanced product candidate, CITY-FXI, is a City-engineered GalNAc-conjugated siRNA therapeutic targeting FXI for the prevention and treatment of thrombosis. CITY-FXI is currently in a Phase 1 clinical trial. Our second product candidate, CITY-RBP4, is a City-engineered GalNAc-conjugated siRNA therapeutic, in development for the treatment of Stargardt disease, which is the most common inherited macular degenerative disease in children and young adults, and other ocular diseases. In mid-2026, we filed an application for CITY-RBP4 to initiate a Phase 1 clinical trial in late 2026. Our third program, CITY-TFR2, is a City-engineered GalNAc-conjugated siRNA therapeutic in development for the treatment of anemia of chronic disease. An IIT for CITY-TFR2 is expected to begin in China for the treatment of anemia in myelofibrosis patients in late 2026, and we plan to file an IND or IND-equivalent application in late 2026 and initiate a Phase 1/2 study for CITY-TFR2 in early 2027. In addition to our wholly owned pipeline, we have entered into research collaboration and license agreements with two industry collaboration partners: in July 2024, we entered into the B+L Agreement with Bausch + Lomb, focused on the discovery and development of intravitreally administered RNAi therapeutics in ocular diseases; and in May 2025, we entered into the Biogen Agreement with Biogen, focused on the discovery and development of RNAi triggers for use in systemic delivery of RNAi to treat neurodegenerative conditions. In addition, we entered into the OSIF License Agreement with OSIF in October 2023, pursuant to which OSIF granted us an exclusive, royalty-bearing license to certain patent rights relating to the originating technology underlying cityRNAs and certain other aspects of our technology platform.
We were incorporated in the State of Delaware in July 2023. We have our principal executive offices located in Cambridge, Massachusetts. Our wholly owned subsidiary, City Therapeutics Securities Corporation, is a Massachusetts securities corporation.
We have devoted substantially all of our efforts to research and development activities, advancing development of the portfolio of programs and platforms, raising capital, developing and maintaining our intellectual property, hiring personnel, and providing general and administrative support for these operations. Since inception, we have funded our operations with proceeds from the issuance and sale of convertible preferred stock, including $4.5 million in gross proceeds from sales of our Series Seed convertible preferred stock from September 2023 through March 2024, $134.7 million in gross
proceeds from sales of our Series A convertible preferred stock from April 2024 through December 2024, and $99.5 million in gross proceeds from sales of our Series B convertible preferred stock from May 2026 through June 2026, the issuance of convertible notes in the amounts of $10.0 million and $30.0 million to Bausch + Lomb in July 2024 and Biogen in May 2025, respectively, and from non-refundable upfront payments of $5.0 million and $16.0 million from our collaboration arrangements with Bausch + Lomb in July 2024 and Biogen in June 2025, respectively. As of June 30, 2026, we had cash, cash equivalents, and restricted cash of $172.1 million.
We have incurred significant operating losses since our inception and expect to continue to generate operating losses for the foreseeable future. Our ability to generate revenue from product sales sufficient to achieve profitability will depend on the successful development and eventual commercialization of any product candidates we may develop. For the years ended December 31, 2024 and 2025, we had a net loss of $27.8 million and $58.5 million, respectively, and $24.3 million and $53.4 million for the six months ended June 30, 2025 and 2026, respectively. As of December 31, 2025 and June 30, 2026, we had an accumulated deficit of $89.1 million and $142.5 million, respectively. We expect our expenses and operating losses will increase substantially as we:
•
advance our product candidates through preclinical and clinical development, including continuing the Phase 1 clinical trial for CITY-FXI and advancing other programs through preclinical and clinical development;
•
seek regulatory approvals for our product candidates that successfully complete clinical trials from Regulatory Authorities;
•
hire additional clinical, quality control, medical, scientific, and other technical personnel to support the clinical development of our product candidates;
•
expand our internal capabilities to support clinical-stage development;
•
increase our headcount as we expand our research and development organization and pre-commercial planning activities;
•
undertake any activities to establish sales, marketing, and distribution capabilities in preparation for commercialization, as applicable, including hiring additional personnel to support such operations;
•
seek to identify, acquire, and develop additional product candidates where we can leverage our City Platform, including through business development efforts to invest in or in-license other technologies or product candidates;
•
maintain, expand, and protect our intellectual property portfolio;
•
experience heightened regulatory scrutiny;
•
make milestone, royalty, or other payments due under the OSIF License Agreement, and any future in-license or collaboration agreements with third parties; and
•
incur additional legal, audit, regulatory, tax and other expenses associated with operating as a public company.
We do not expect to generate any revenue from product sales unless and until we successfully complete the clinical development or future clinical development of, and obtain regulatory approval for, one or more of our current or future product candidates, which may not occur for several years, if at all. In addition, if we obtain marketing approval for our product candidates and any other product candidates we may identify and pursue, we expect to incur significant commercialization expenses related to product manufacturing, sales, marketing and distribution.
Our net losses may fluctuate significantly from quarter-to-quarter and year-to-year, depending on the timing of our preclinical studies and planned clinical trials and our expenditures on other research and development activities. There can be no assurance that our research and development efforts will be successfully completed, that adequate protection for our intellectual property will be obtained, that any products developed will obtain necessary government regulatory approval or that any approved products
will be commercially viable. Even if our product development efforts are successful, it is uncertain when, if ever, we will generate significant revenue from product sales. We operate in an environment of rapid change in technology and substantial competition from pharmaceutical and biotechnology companies. In addition, we are dependent upon the services of our employees and consultants. Accordingly, until such time as we can generate significant revenue from our existing or future product candidates, if ever, we expect to finance our cash needs through private and public equity financings, additional collaborations, strategic alliances and marketing, distribution or license arrangements. See the section titled “Liquidity and Capital Resources.” However, we may be unable to obtain financing on acceptable terms, or at all, and we may not be able to enter into additional collaborations or other arrangements. The terms of any financing may adversely affect the holdings or the rights of our stockholders. Our inability to raise capital or enter into such other arrangements when needed would have a negative impact on our financial condition and could force us to delay, scale back or discontinue some or all of our research and development programs, product portfolio expansion efforts or commercialization efforts. Although we continue to pursue these plans, there is no assurance that we will be successful in obtaining sufficient funding on terms acceptable to us to fund continuing operations, if at all.
We believe, based on our current operating plan, that our cash and cash equivalents as of June 30, 2026, together with the net proceeds from this offering, will be sufficient to fund our operating expenses and capital expenditure requirements into late 2028. See sections titled “Use of Proceeds,” “Liquidity and Capital Resources,” “Risk Factors — Risks Related to Our Limited Operating History, Financial Condition and Need for Additional Capital,” and Note 1 to our audited consolidated financial statements.
Components of Results of Operations
The following discussion summarizes the key components of our results of operations.
Collaboration Revenue
Our revenue to date has been generated primarily from our research collaboration and license agreements with Bausch + Lomb and Biogen. We recognize revenue under these arrangements over time as we perform the research services using an input method which is based on the actual costs incurred relative to total costs expected to be incurred. We have not generated any revenue from product sales and do not expect to do so for the foreseeable future. We expect that any revenue we generate from our existing collaborations will fluctuate from period to period based on the timing of our research and development activities, and our ability to achieve specified milestones. If we enter into additional collaboration or license agreements with third parties, we may generate revenue in the future from payments from such collaboration or license agreements, or any combination thereof. However, there can be no assurance as to when we will generate such revenue, if at all.
Operating Expenses
Research and Development Expenses
Research and development costs are expensed as incurred. Research and development costs include salaries, stock-based compensation and other employee benefit expenses, consultants, costs of funding research performed by third parties on our behalf, lab-related supplies, and allocated facility-related expenses. We do not track, record or maintain personnel-related expenses or other indirect or shared operating costs incurred for our research and development programs on a program-by-program basis, because our personnel and other resources are deployed across multiple product candidates and research and development programs and, as a result, these costs are not separately identifiable by program.
Non-refundable advance payments for goods and services to be received in the future for use in research and development activities are recorded as prepaid expenses on the accompanying
consolidated balance sheets. The prepaid amounts are expensed as the related goods are delivered or the services are performed, or when it is no longer expected that the goods will be delivered, or the services rendered.
We record accrued liabilities for estimated costs of research and development activities conducted by third-party service providers. We accrue these costs based on factors such as estimates of the work completed and in accordance with agreements established with our third-party service providers under the service agreements. We make significant judgments and estimates in determining the accrued liabilities balance in each reporting period. As actual costs become known, we adjust our accrued liabilities. The historical accrual estimates made by us have not been materially different from the actual costs.
We expect research and development expenses to increase as we continue advancing our current and future product candidates through preclinical and clinical development and invest in the development of additional product candidates.
General and Administrative Expenses
General and administrative expenses consist primarily of salaries and personnel-related costs, including stock-based compensation expense, for our personnel in executive, business development, legal, finance and accounting, human resources and other administrative functions. General and administrative expenses also include consulting fees, facility costs not otherwise included in research and development expenses, fees paid for accounting and tax services, insurance expenses, and legal costs consisting of general corporate legal fees and patent legal fees.
We anticipate that our general and administrative expenses will increase in the future as we increase our headcount to support the expansion of our business, particularly in support of development of product candidates and our continued research and clinical development activities. We will also incur significant costs associated with being a public company, including increased accounting, audit, legal, regulatory, compliance, and director and officer insurance costs, as well as expenses related to services associated with maintaining compliance with the requirements of the Nasdaq Stock Market, the SEC, and investor relations costs.
Total Other Income (Expense), Net
Total other income (expense), net, consists primarily of (i) interest income earned on our cash and cash equivalents, (ii) interest expense on our convertible notes (the “Original Notes”) issued to ARCH Venture Fund XII L.P. (“ARCH”) and one of our founders (the “Founder”), as well as the convertible note issued to Bausch + Lomb (“B+L Note”) and the Biogen Note in connection with our research collaboration and license agreements, (iii) the changes in the fair value of these convertible notes recorded through earnings as they were accounted for under the fair value option, and (iv) gain resulting from our lease modification.
Results of Operations
Comparison of the years ended December 31, 2024 and 2025
The following table summarizes our results of operations (in thousands):
| |
|
|
For the year ended
December 31,
|
|
|
|
|
| |
|
|
2024
|
|
|
2025
|
|
|
Change
|
|
|
Collaboration revenue
|
|
|
|
$ |
2,778 |
|
|
|
|
$ |
13,562 |
|
|
|
|
$ |
10,784 |
|
|
|
Operating expenses:
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
Research and development
|
|
|
|
|
25,682 |
|
|
|
|
|
58,420 |
|
|
|
|
|
32,738 |
|
|
|
General and administrative
|
|
|
|
|
6,186 |
|
|
|
|
|
15,064 |
|
|
|
|
|
8,878 |
|
|
|
Total operating expenses
|
|
|
|
|
31,868 |
|
|
|
|
|
73,484 |
|
|
|
|
|
41,616 |
|
|
|
Loss from operations
|
|
|
|
|
(29,090) |
|
|
|
|
|
(59,922) |
|
|
|
|
|
(30,832) |
|
|
| |
|
|
For the year ended
December 31,
|
|
|
|
|
| |
|
|
2024
|
|
|
2025
|
|
|
Change
|
|
|
Other income:
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
Interest income
|
|
|
|
|
2,397 |
|
|
|
|
|
4,208 |
|
|
|
|
|
1,811 |
|
|
|
Interest expense
|
|
|
|
|
(448) |
|
|
|
|
|
(2,257) |
|
|
|
|
|
(1,809) |
|
|
|
Other expense, net
|
|
|
|
|
(695) |
|
|
|
|
|
(534) |
|
|
|
|
|
161 |
|
|
|
Total other income, net
|
|
|
|
|
1,254 |
|
|
|
|
|
1,417 |
|
|
|
|
|
163 |
|
|
|
Net loss
|
|
|
|
$ |
(27,836) |
|
|
|
|
$ |
(58,505) |
|
|
|
|
$ |
(30,669) |
|
|
| |
Collaboration Revenue
The following table summarizes our collaboration revenues (in thousands):
| |
|
|
For the year ended
December 31,
|
|
|
|
|
| |
|
|
2024
|
|
|
2025
|
|
|
Change
|
|
|
Revenue recognized from B+L Agreement
|
|
|
|
$ |
2,778 |
|
|
|
|
$ |
7,123 |
|
|
|
|
$ |
4,345 |
|
|
|
Revenue recognized from Biogen Agreement
|
|
|
|
|
— |
|
|
|
|
|
6,439 |
|
|
|
|
|
6,439 |
|
|
|
Total Revenue
|
|
|
|
$ |
2,778 |
|
|
|
|
$ |
13,562 |
|
|
|
|
$ |
10,784 |
|
|
Our collaboration revenue increased by $10.8 million, from $2.8 million for the year ended December 31, 2024 to $13.6 million for the year ended December 31, 2025. The increase was attributed to $6.4 million from the Biogen Agreement, which was executed during 2025, and an increase of $4.3 million pursuant to the B+L Agreement as we continued to perform research and development activities since its execution in July 2024.
Research and Development Expenses
The following table summarizes our research and development expenses (in thousands):
| |
|
|
For the year ended
December 31,
|
|
|
|
|
| |
|
|
2024
|
|
|
2025
|
|
|
Change
|
|
|
Direct costs:
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
CITY-FXI
|
|
|
|
$ |
5,694 |
|
|
|
|
$ |
12,161 |
|
|
|
|
$ |
6,467 |
|
|
|
CITY-RBP4
|
|
|
|
|
2,981 |
|
|
|
|
|
7,109 |
|
|
|
|
|
4,128 |
|
|
|
CITY-TFR2
|
|
|
|
|
1,104 |
|
|
|
|
|
2,985 |
|
|
|
|
|
1,881 |
|
|
|
Ocular
|
|
|
|
|
919 |
|
|
|
|
|
2,563 |
|
|
|
|
|
1,644 |
|
|
|
CNS
|
|
|
|
|
240 |
|
|
|
|
|
2,711 |
|
|
|
|
|
2,471 |
|
|
|
Indirect costs:
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
Personnel-related expenses (excluding stock-based compensation)
|
|
|
|
|
4,938 |
|
|
|
|
|
10,498 |
|
|
|
|
|
5,560 |
|
|
|
Stock-based compensation
|
|
|
|
|
445 |
|
|
|
|
|
955 |
|
|
|
|
|
510 |
|
|
|
Other indirect research and development
|
|
|
|
|
9,361 |
|
|
|
|
|
19,438 |
|
|
|
|
|
10,077 |
|
|
|
Total research and development expenses
|
|
|
|
$ |
25,682 |
|
|
|
|
$ |
58,420 |
|
|
|
|
$ |
32,738 |
|
|
Research and development expenses increased by $32.7 million, from $25.7 million for the year ended December 31, 2024 to $58.4 million for the year ended December 31, 2025. The increase in research and development expenses reflects continued advancement of multiple pipeline programs from discovery into IND- and IND-equivalent-enabling development and clinical-stage activities, expansion of our collaboration programs, and continued investment in our platform and research infrastructure, including:
•
$6.5 million of increased costs associated with our lead program, CITY-FXI, which was primarily due to IND or IND-equivalent and manufacturing costs such as CMC activities, drug substance and drug product manufacturing, as well as preclinical and toxicology studies and clinical readiness costs ahead of the filing of the IND or IND-equivalent application;
•
$4.1 million of increased costs associated with CITY-RBP4, which was attributed to increased preclinical and IND or IND-equivalent-enabling activities such as preclinical pharmacology and toxicology studies, CMC activities, drug-metabolism and pharmacokinetics studies, as we advanced CITY-RBP4 toward a planned IND or IND-equivalent application filing;
•
$1.9 million of increased costs associated with our CITY-TFR2 program, which was primarily due to increased preclinical research and discovery activities as we advanced our CITY-TFR2 program;
•
$1.6 million of increased costs associated with research activities performed under the B+L Agreement, which was executed in July 2024;
•
$2.5 million of increased costs associated with research activities performed under the Biogen Agreement, which was executed in May 2025;
•
$5.6 million of increased employee compensation expense (excluding stock-based compensation) primarily due to increased research and development headcount to support our growing pipeline and platform activities;
•
$0.5 million of increased stock-based compensation expense primarily due to additional equity awards granted under our 2023 Plan; and
•
$10.1 million of increased costs associated with other indirect research and development costs, which primarily included (i) a $2.9 million increase in facilities and depreciation costs associated with the expanded lease for our Cambridge, Massachusetts office and laboratory space, (ii) a $2.3 million increase in equipment and laboratory supplies, (iii) a $2.3 million increase in overhead and indirect costs to support our growing platform, pipeline and research and development activities, and (iv) a $2.0 million increase in outside research and consulting costs.
General and Administrative Expenses
The following table summarizes our general and administrative expenses (in thousands):
| |
|
|
For the year ended
December 31,
|
|
|
|
|
| |
|
|
2024
|
|
|
2025
|
|
|
Change
|
|
|
Employee compensation (excluding stock-based compensation)
|
|
|
|
$ |
1,197 |
|
|
|
|
$ |
3,907 |
|
|
|
|
$ |
2,710 |
|
|
|
Stock-based compensation
|
|
|
|
|
560 |
|
|
|
|
|
2,042 |
|
|
|
|
|
1,482 |
|
|
|
Other general and administrative expenses
|
|
|
|
|
4,429 |
|
|
|
|
|
9,115 |
|
|
|
|
|
4,686 |
|
|
|
Total general and administrative expenses
|
|
|
|
$ |
6,186 |
|
|
|
|
$ |
15,064 |
|
|
|
|
$ |
8,878 |
|
|
General and administrative expenses increased by $8.9 million, from $6.2 million for the year ended December 31, 2024 to $15.1 million for the year ended December 31, 2025. The increase in general and administrative expenses was primarily attributable to:
•
$2.7 million of increased employee compensation expense (excluding stock-based compensation) primarily due to increased general and administrative headcount to support our operations;
•
$1.5 million of increased stock-based compensation expense primarily due to additional equity awards granted under our 2023 Plan; and
•
$4.7 million of increased other general and administrative expenses primarily due to increased professional fees, facilities, depreciation, information technology and other general and administrative costs that are not personnel-related.
Total Other Income, Net
Total other income, net increased by $0.2 million, from $1.2 million for the year ended December 31, 2024 to $1.4 million for the year ended December 31, 2025. Interest income increased by $1.8 million primarily due to higher cash and cash equivalents balances following the closing of additional Series A convertible preferred stock issuances. This increase was substantially offset by an increase of $1.8 million in interest expense recognized on our Biogen Note and B+L Note, which was accounted for under the fair value option. Other expense, net, decreased by $0.2 million, which included changes in the fair value of such convertible notes recorded through earnings and offset by the gain recorded for our lease modification.
Comparison of the six months ended June 30, 2025 and 2026
The following table summarizes our results of operations (in thousands):
| |
|
|
For the six months
ended June 30,
|
|
|
Change
|
|
| |
|
|
2025
|
|
|
2026
|
|
|
Collaboration revenue
|
|
|
|
$ |
5,578 |
|
|
|
|
$ |
6,358 |
|
|
|
|
$ |
780 |
|
|
|
Operating expenses:
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
Research and development
|
|
|
|
|
24,375 |
|
|
|
|
|
46,319 |
|
|
|
|
|
21,944 |
|
|
|
General and administrative
|
|
|
|
|
7,796 |
|
|
|
|
|
11,170 |
|
|
|
|
|
3,374 |
|
|
|
Total operating expenses
|
|
|
|
|
32,171 |
|
|
|
|
|
57,489 |
|
|
|
|
|
25,318 |
|
|
|
Loss from operations
|
|
|
|
|
(26,593) |
|
|
|
|
|
(51,131) |
|
|
|
|
|
(24,538) |
|
|
|
Other income (expense), net:
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
Interest income
|
|
|
|
|
2,064 |
|
|
|
|
|
1,595 |
|
|
|
|
|
(469) |
|
|
|
Interest expense
|
|
|
|
|
(646) |
|
|
|
|
|
(1,496) |
|
|
|
|
|
(850) |
|
|
|
Other income (expense), net
|
|
|
|
|
893 |
|
|
|
|
|
(2,411) |
|
|
|
|
|
(3,304) |
|
|
|
Total other income (expense), net
|
|
|
|
|
2,311 |
|
|
|
|
|
(2,312) |
|
|
|
|
|
(4,623) |
|
|
|
Net loss
|
|
|
|
$ |
(24,282) |
|
|
|
|
$ |
(53,443) |
|
|
|
|
$ |
(29,161) |
|
|
Collaboration Revenue
The following table summarizes our collaboration revenues (in thousands):
| |
|
|
For the six months
ended June 30,
|
|
|
Change
|
|
| |
|
|
2025
|
|
|
2026
|
|
|
Revenue recognized from B+L Agreement
|
|
|
|
$ |
3,567 |
|
|
|
|
$ |
2,845 |
|
|
|
|
$ |
(722) |
|
|
|
Revenue recognized from Biogen Agreement
|
|
|
|
|
2,011 |
|
|
|
|
|
3,513 |
|
|
|
|
|
1,502 |
|
|
|
Total revenue
|
|
|
|
$ |
5,578 |
|
|
|
|
$ |
6,358 |
|
|
|
|
$ |
780 |
|
|
Our collaboration revenue increased by $0.8 million, from $5.6 million for the six months ended June 30, 2025 to $6.4 million for the six months ended June 30, 2026. The increase was attributed to $1.5 million from the Biogen Agreement, which was executed during 2025, partially offset by a decrease of $0.7 million pursuant to the B+L Agreement, reflecting the timing and level of research and development activities performed under the agreement since its execution in July 2024.
Research and Development Expenses
The following table summarizes our research and development expenses (in thousands):
| |
|
|
For the six months
ended June 30,
|
|
|
Change
|
|
| |
|
|
2025
|
|
|
2026
|
|
|
Direct costs:
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
CITY-FXI
|
|
|
|
$ |
6,217 |
|
|
|
|
$ |
5,565 |
|
|
|
|
$ |
(652) |
|
|
|
CITY-RBP4
|
|
|
|
|
2,519 |
|
|
|
|
|
6,337 |
|
|
|
|
|
3,818 |
|
|
|
CITY-TFR2
|
|
|
|
|
516 |
|
|
|
|
|
3,543 |
|
|
|
|
|
3,027 |
|
|
|
Ocular
|
|
|
|
|
1,053 |
|
|
|
|
|
734 |
|
|
|
|
|
(319) |
|
|
|
CNS
|
|
|
|
|
1,115 |
|
|
|
|
|
1,182 |
|
|
|
|
|
67 |
|
|
|
Undisclosed programs
|
|
|
|
|
— |
|
|
|
|
|
1,409 |
|
|
|
|
|
1,409 |
|
|
|
Indirect costs:
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
Personnel-related expenses (excluding stock-based compensation)
|
|
|
|
|
4,726 |
|
|
|
|
|
9,755 |
|
|
|
|
|
5,029 |
|
|
|
Stock-based compensation
|
|
|
|
|
442 |
|
|
|
|
|
1,316 |
|
|
|
|
|
874 |
|
|
|
Other indirect research and development
|
|
|
|
|
7,787 |
|
|
|
|
|
16,478 |
|
|
|
|
|
8,691 |
|
|
|
Total research and development expenses
|
|
|
|
$ |
24,375 |
|
|
|
|
$ |
46,319 |
|
|
|
|
$ |
21,944 |
|
|
Research and development expenses increased by $21.9 million, from $24.4 million for the six months ended June 30, 2025 to $46.3 million for the six months ended June 30, 2026. The increase in research and development expenses reflects continued advancement of multiple pipeline programs from discovery into IND- and IND-equivalent-enabling development and clinical-stage activities, expansion of our collaboration programs, and continued investment in our platform and research infrastructure, including:
•
$0.7 million of decreased costs associated with our lead program, CITY-FXI, which was primarily due to the substantial completion in the 2025 period of IND or IND-equivalent-enabling activities, including CMC, drug substance and drug product manufacturing, and preclinical and toxicology studies, partially offset by increased clinical trial costs following the initiation of our Phase 1 clinical trial in the 2026 period; $3.8 million of increased costs associated with CITY-RBP4, which was attributed to increased preclinical and IND or IND-equivalent-enabling activities such as preclinical pharmacology and toxicology studies, CMC activities, drug-metabolism and pharmacokinetics studies, as we advanced CITY-RBP4 toward a planned IND or IND-equivalent application filing;
•
$3.0 million of increased costs associated with our CITY-TFR2 program, which was primarily due to increased preclinical research and discovery activities as we advanced our CITY-TFR2 program;
•
$0.3 million of decreased costs associated with research activities performed under the B+L Agreement in accordance with the research plan and candidate selection activities;
•
$0.1 million of increased costs associated with research activities performed under the Biogen Agreement due to continued spend on research activities;
•
$1.4 million of increased costs associated with our undisclosed programs, reflecting new discovery-stage activity initiated during 2026;
•
$5.0 million of increased employee compensation expense (excluding stock-based compensation) primarily due to increased research and development headcount to support our growing pipeline and platform activities;
•
$0.9 million of increased stock-based compensation expense primarily due to additional equity awards granted under our 2023 Plan and achievement of a performance-based milestone; and
•
$8.7 million of increased costs associated with other indirect research and development costs, which primarily included (i) a $1.5 million increase in facilities and depreciation costs associated with the expanded lease for our Cambridge, Massachusetts office and laboratory space, (ii) a $4.7 million increase in overhead and indirect costs to support our growing platform, pipeline and research and development activities, (iii) a $0.6 million increase in outside research and consulting costs, and (iv) a $1.9 million increase in platform research costs.
General and Administrative Expenses
The following table summarizes our general and administrative expenses (in thousands):
| |
|
|
For the six months
ended June 30,
|
|
|
Change
|
|
| |
|
|
2025
|
|
|
2026
|
|
|
Employee compensation (excluding stock-based compensation)
|
|
|
|
$ |
2,119 |
|
|
|
|
$ |
2,199 |
|
|
|
|
$ |
80 |
|
|
|
Stock-based compensation
|
|
|
|
|
1,055 |
|
|
|
|
|
3,601 |
|
|
|
|
|
2,546 |
|
|
|
Other general and administrative expenses
|
|
|
|
|
4,622 |
|
|
|
|
|
5,370 |
|
|
|
|
|
748 |
|
|
|
Total general and administrative expenses
|
|
|
|
$ |
7,796 |
|
|
|
|
$ |
11,170 |
|
|
|
|
$ |
3,374 |
|
|
General and administrative expenses increased by $3.4 million, from $7.8 million for the six months ended June 30, 2025 to $11.2 million for the six months ended June 30, 2026. The increase in general and administrative expenses was primarily attributable to:
•
$0.1 million of increased employee compensation expense (excluding stock-based compensation) primarily due to increased general and administrative headcount to support our operations;
•
$2.5 million of increased stock-based compensation expense primarily due to additional equity awards granted under our 2023 Plan and the achievement of a performance-based milestone; and
•
$0.7 million of increased other general and administrative expenses primarily due to increased professional fees, facilities, depreciation, information technology and other general and administrative costs that are not personnel-related.
Total Other Income (Expense), Net
Total other income (expense), net decreased by $4.6 million, from income of $2.3 million for the six months ended June 30, 2025 to expense of $2.3 million for the six months ended June 30, 2026. Interest income decreased by $0.5 million primarily due to lower average interest-earning cash balances during the 2026 period. This decrease was substantially compounded by an increase of $0.9 million in interest expense, reflecting a full six months of interest on the $30.0 million Biogen Note issued in May 2025 compared with approximately one month in the 2025 period, together with interest on the $10.0 million B+L Note through its conversion in May 2026. Other income (expense), net, decreased by $3.3 million, from income of $0.9 million to expense of $2.4 million, primarily due to a $2.2 million increase in expense from changes in the fair value of our convertible notes, which are accounted for under the fair value option, and the recognition of a $1.0 million gain related to the modification of our lease in 2025.
Liquidity and Capital Resources
Sources of Liquidity
We have incurred net losses and negative cash flows from operations since our inception and anticipate we will continue to incur net losses for the foreseeable future.
Since our inception in July 2023, we have funded our operations primarily through (i) proceeds from sales of our convertible preferred stock, (ii) proceeds from issuances of convertible notes to our
collaboration partners, and (iii) upfront and milestone payments under our research collaboration and license agreements with Bausch + Lomb and Biogen. As of June 30, 2026, we had cash and cash equivalents of $169.6 million, restricted cash of $2.5 million, and an accumulated deficit of $142.5 million.
We have issued both convertible notes and convertible preferred stock, which are classified as a liability and as mezzanine equity, respectively, on our consolidated balance sheets. Immediately prior to the closing of this offering, (i) all outstanding shares of our convertible preferred stock will automatically convert into common stock on a one-for-one basis, and (ii) the outstanding principal and accrued interest under the Biogen Note will automatically convert into common stock. As a result, no shares of convertible preferred stock and no convertible notes will remain outstanding after this offering, the carrying value of our convertible preferred stock will be reclassified from mezzanine equity to permanent stockholders’ equity, and the Biogen Note will be reclassified from a liability to permanent stockholders’ equity. For a description of the material terms of our convertible note and convertible preferred stock, see Notes 5 and 10, respectively, to our audited consolidated financial statements and to our unaudited interim condensed consolidated financial statements.
Cash Flows
Comparison of the years ended December 31, 2024 and 2025
The following table provides information regarding our cash flows for the periods presented (in thousands):
| |
|
|
For the year ended
December 31,
|
|
| |
|
|
2024
|
|
|
2025
|
|
|
Net cash used in operating activities
|
|
|
|
$ |
(20,391) |
|
|
|
|
$ |
(41,611) |
|
|
|
Net cash used in investing activities
|
|
|
|
|
(3,379) |
|
|
|
|
|
(2,378) |
|
|
|
Net cash provided by financing activities
|
|
|
|
|
149,438 |
|
|
|
|
|
30,859 |
|
|
|
Net increase (decrease) in cash, cash equivalents, and restricted cash(1)
|
|
|
|
$ |
125,668 |
|
|
|
|
$ |
(13,130) |
|
|
(1)
Cash, cash equivalents, and restricted cash includes restricted cash of $1.5 million and $2.5 million as of December 31, 2024 and 2025, respectively.
Net Cash Used in Operating Activities
Net cash used in operating activities was $20.4 million for the year ended December 31, 2024, compared to $41.6 million for the year ended December 31, 2025. The increase in cash used in operating activities was primarily attributable to our higher net loss, which increased by $30.7 million year-over-year, principally driven by increased research and development activities and headcount, and increases in prepaid expenses and other current assets, due to timing. This increase was partially offset by changes in our operating assets and liabilities, including an increase in deferred revenue from our collaboration arrangements driven by timing.
Net Cash Used in Investing Activities
Net cash used in investing activities was $3.4 million for the year ended December 31, 2024, compared to $2.4 million for the year ended December 31, 2025. In both periods, cash used in investing activities consisted primarily of purchases of property and equipment, laboratory equipment and leasehold improvements at our Cambridge, Massachusetts facilities, partially offset by minimal proceeds from sales of property and equipment. The decrease of $1.0 million was primarily attributable to a $1.1 million reduction in purchases of property and equipment.
Net Cash Provided by Financing Activities
Net cash provided by financing activities was $149.4 million for the year ended December 31, 2024, primarily consisting of $134.7 million in gross proceeds from the issuance of our Series A
convertible preferred stock and $15.0 million in gross proceeds from issuances of convertible notes (including the B+L Note and the Original Notes that converted into Series A convertible preferred stock), partially offset by $0.3 million in offering costs. Net cash provided by financing activities was $30.9 million for the year ended December 31, 2025, primarily consisting of $30.0 million in gross proceeds from the issuance of the Biogen Note in May 2025 and $0.9 million in proceeds from exercises of stock options.
Comparison of the six months ended June 30, 2025 and 2026
The following table provides information regarding our cash flows for the periods presented (in thousands):
| |
|
|
For the six months
ended June 30,
|
|
| |
|
|
2025
|
|
|
2026
|
|
|
Net cash used in operating activities
|
|
|
|
$ |
(7,098) |
|
|
|
|
$ |
(43,158) |
|
|
|
Net cash used in investing activities
|
|
|
|
|
(352) |
|
|
|
|
|
(1,022) |
|
|
|
Net cash provided by financing activities
|
|
|
|
|
30,806 |
|
|
|
|
|
100,992 |
|
|
|
Net increase in cash, cash equivalents, and restricted cash(1)
|
|
|
|
$ |
23,356 |
|
|
|
|
$ |
56,812 |
|
|
(1)
Cash, cash equivalents, and restricted cash includes restricted cash of $2.5 million as of each of June 30, 2025 and 2026.
Net Cash Used in Operating Activities
Net cash used in operating activities was $7.1 million for the six months ended June 30, 2025, compared to $43.2 million for the six months ended June 30, 2026. The increase in cash used in operating activities was primarily attributable to our higher net loss, which increased by $29.2 million period-over-period, principally driven by increased expenses from research and development activities and headcount. This increase was further impacted by changes in our operating assets and liabilities, including a decrease in deferred revenue from our collaboration arrangements and an increase in accounts receivable due to timing.
Net Cash Used in Investing Activities
Net cash used in investing activities was $0.4 million for the six months ended June 30, 2025, compared to $1.0 million for the six months ended June 30, 2026. In both periods, cash used in investing activities consisted primarily of purchases of property and equipment, laboratory equipment and leasehold improvements at our Cambridge, Massachusetts facilities. The increase of $0.7 million was primarily attributable to increased purchases of property and equipment.
Net Cash Provided by Financing Activities
Net cash provided by financing activities was $30.8 million for the six months ended June 30, 2025, primarily consisting of $30.0 million in gross proceeds from the issuance of the Biogen Note and $0.8 million in proceeds from exercises of stock options. Net cash provided by financing activities was $101.0 million for the six months ended June 30, 2026, primarily consisting of $99.5 million in gross proceeds from the issuance of Series B convertible preferred stock and $2.1 million in proceeds from exercises of stock options.
Future Funding Requirements
As of June 30, 2026, we had cash, cash equivalents, and restricted cash of $172.1 million.
We expect that our cash and cash equivalents as of June 30, 2026 will be sufficient to fund our operating expenses and capital expenditure requirements through at least 12 months from the date our unaudited interim condensed consolidated financial statements were available to be issued.
We believe, based on our current operating plan, that the net proceeds from this offering, together with our existing cash and cash equivalents, will enable us to fund our planned operating expenses and capital expenditure requirements into late 2028.
Due to the inherently unpredictable nature of preclinical and clinical development and given the early stage of our programs and product candidates, we cannot reasonably estimate the costs we will incur and the timelines that will be required to complete development, obtain any marketing approval, and commercialize our products, if and when approved. For the same reasons, we are also unable to predict when, if ever, we will generate revenue from product sales or whether, or when, if ever, we may achieve profitability. Clinical and preclinical development timelines, the probability of success, and development costs can differ materially from expectations. In addition, we cannot forecast which products, if approved, may be subject to future collaborations, when such arrangements will be secured, if at all, and to what degree such arrangements would affect our development plans and capital requirements. We will need to raise substantial additional capital in the future.
Our primary uses of capital are to fund research and development activities, compensation and related expenses, and general overhead costs. We expect our expenses to increase substantially in connection with our ongoing activities, particularly as we advance our current and future product candidates through discovery, preclinical studies, and clinical trials.
Our funding requirements and timing and amount of our operating expenditures will depend on many factors, including:
•
the scope, timing, progress, costs, and results of discovery, preclinical development and clinical trials for our current or future product candidates and effectiveness of our platform;
•
the number of clinical trials required for regulatory approval of our current or future product candidates;
•
the costs, timing, and outcome of regulatory review of any of our current or future product candidates;
•
the costs associated with acquiring or licensing additional product candidates, technologies or assets, including the timing and amount of any milestones, royalties, or other payments due in connection with our acquisitions and licenses, as applicable;
•
the cost of manufacturing clinical and commercial supplies of our current or future product candidates;
•
the costs and timing of preparing, filing, and prosecuting patent applications, maintaining and enforcing our intellectual property rights and defending any intellectual property-related claims, including any claims by third parties that we are infringing upon their intellectual property rights;
•
our ability to establish new, and maintain existing, strategic collaborations, licensing or other arrangements on favorable terms, if at all, and the financial terms of any such agreements, including the timing and amount of any future milestone, royalty or other payments due to or payable by us thereunder and the extent to which we are obligated to reimburse, or entitled to reimbursement of, clinical trial costs under any such agreements;
•
the costs and timing of future commercialization activities, including manufacturing, marketing, sales, and distribution, for any of our product candidates for which we receive marketing approval;
•
the revenue, if any, received from commercial sales of our product candidates for which we receive marketing approval;
•
expenses to attract, hire, and retain skilled personnel;
•
the costs of operating as a public company;
•
our ability to establish a commercially viable pricing structure and obtain approval for coverage and adequate reimbursement from third-party and government payors;
•
the effect of macroeconomic trends including inflation, tariffs, and fluctuating interest rates;
•
addressing any potential supply chain interruptions or delays;
•
the effect of competing technological and market developments; and
•
the extent to which we acquire or invest in businesses, products, and technologies.
Until such time, if ever, that we can generate substantial product revenue, we expect to finance our operations through a combination of public and private equity offerings, debt and royalty financings, or other sources of capital, which may include additional collaborations with other companies, or licensing arrangements with third parties, or other similar arrangements. However, we may be unable to raise additional funds or enter into such other arrangements when needed on favorable terms or at all. To the extent that we raise additional capital through the sale of equity or convertible debt securities, the ownership interest of our stockholders will or could be diluted, and the terms of these securities may include liquidation or other preferences that adversely affect the rights of our common stockholders. Debt or royalty financing and preferred equity financing, if available, may involve agreements that include covenants limiting or restricting our ability to take specific actions, such as incurring additional debt, making acquisitions or capital expenditures, or declaring dividends. If we raise additional funds through additional collaborations, strategic alliances, or licensing arrangements with third parties, we may have to relinquish valuable rights to our technologies, future revenue streams, research programs, product candidates, or grant licenses on terms that may not be favorable to us. If we are unable to raise additional capital or obtain adequate funding when needed or on acceptable terms, we may be required to delay, scale back, or discontinue our research, product development, or future commercialization efforts, or grant rights to develop and market product candidates that we would otherwise prefer to develop and market ourselves.
License and Collaboration Agreements
Below is a summary of the key terms for our material license and collaboration agreements. For a more detailed description, see the section of this prospectus titled “Business — License and Collaboration Agreements,” and Note 9 to our audited consolidated financial statements.
Bausch + Lomb Research Collaboration and License Agreement
On July 8, 2024, we entered into the B+L Agreement with Bausch + Lomb, pursuant to which (i) we agreed to undertake a research program to discover certain licensed molecules which would enable Bausch + Lomb to develop and subsequently commercialize, a licensed product in the field of ocular disease and (ii) we granted Bausch + Lomb an option to obtain an exclusive, worldwide license to certain of our intellectual property to develop, manufacture and commercialize a licensed product in the field of ocular disease. Simultaneously with the execution of the B+L Agreement, we issued a convertible note to Bausch + Lomb in an aggregate principal amount of $10.0 million, bearing interest at 8.0% per annum and maturing on July 8, 2027, unless earlier converted, which subsequently converted into shares of our Series B-1 convertible preferred stock.
Pursuant to the B+L Agreement, we will perform specified research activities pursuant to a research plan and deliver to Bausch + Lomb a development candidate for which Bausch + Lomb may exercise its option to license. We plan to substantially complete the research and development under the B+L Agreement to enable Bausch + Lomb to select a development candidate by late 2026. If Bausch + Lomb exercises its option, it will engage in the development, manufacturing and commercialization of licensed molecules and licensed products from the development candidate.
Under the B+L Agreement, we received a non-refundable, non-creditable upfront payment of $5.0 million from Bausch + Lomb in July 2024. Over the course of the research plan, Bausch + Lomb is required to reimburse us for amounts incurred for full-time employees and other related costs and materials directly associated with the research plan. Additionally, we are eligible to receive (i) a certain development candidate selection fee and certain other development, regulatory, commercial and sales milestones, collectively up to an aggregate of $485.0 million, and (ii) tiered royalties ranging from mid-single digits to low teens on annual net sales of licensed products, subject to customary reductions and an
overall royalty floor. If the agreement terminates and we later successfully commercialize a product candidate using the licensed intellectual property, we are required to pay Bausch + Lomb a low-single digit royalty on future net sales. As of June 30, 2026, no such milestone payments or royalties had become due to us.
Biogen Research Collaboration and License Agreement
On May 23, 2025, we entered into the Biogen Agreement with Biogen, pursuant to which (i) we and Biogen shall collaborate to undertake a research plan to discover and develop certain RNAi triggers that inhibit the expression of a target protein and conjugates of such RNAi triggers and (ii) we granted Biogen an exclusive, worldwide, royalty-bearing license to certain of our intellectual property to develop, manufacture and commercialize licensed compounds and licensed products in the field in the territory. Simultaneously with the execution of the Biogen Agreement, we issued a convertible note to Biogen in an aggregate principal amount of $30.0 million, bearing interest at 8.0% per annum and maturing on July 1, 2027, unless earlier converted.
Pursuant to the Biogen Agreement, we will perform research activities and deliver to Biogen data packages and technology transfers to enable Biogen to select and advance a development candidate, for which Biogen may exercise its rights under the license. We plan to substantially complete the research and development under the Biogen Agreement by late 2026 to enable Biogen to select a development candidate by late 2027. If Biogen selects a development candidate, it will control the development, manufacturing and commercialization of licensed compounds and licensed products. Each party is responsible for its own research costs under the agreement. We also granted Biogen an option, exercisable during a specified period and upon specified terms, to designate one additional target, subject to execution of a separate collaboration and license agreement on substantially the same terms and an additional payment of a specified option exercise fee.
Under the Biogen Agreement, we received a non-refundable, non-creditable upfront payment of $16.0 million from Biogen in June 2025. We are also eligible to receive an additional non-refundable upfront payment contingent upon the satisfaction of certain conditions relating to our upstream license with OSIF. Additionally, we are eligible to receive up to an aggregate of approximately $1.0 billion in research, development, commercial and sales milestones, along with tiered royalties ranging from high-single digits to low teens on annual net sales of licensed products, subject to customary reductions and an overall royalty floor. As of June 30, 2026, no development, commercial or sales milestones or royalties had become due to us. During the six months ended June 30, 2026, we achieved a research milestone resulting in $4.0 million becoming due to us.
Contractual Obligations and Commitments
OSIF License Agreement
On October 30, 2023, we entered into the OSIF License Agreement with OSIF, pursuant to which OSIF granted us an exclusive, royalty-bearing license to certain patent rights relating to the originating technology underlying cityRNAs and certain other aspects of our technology platform to make, have made, use, sell, and import certain licensed products, subject to certain terms and restrictions therein. As consideration, we issued to OSIF 622,801 shares of our common stock, which constituted 5% of our fully diluted capitalization as of the effective date of the agreement, valued at $0.1 million, and agreed to issue additional equity securities to OSIF from time to time as necessary to maintain its pro rata ownership percentage until we have received a specified amount of equity financing proceeds pursuant to certain anti-dilution provisions of the OSIF License Agreement. On April 17, 2024, upon the closing of our Series A convertible preferred stock financing, we issued to OSIF an additional 32,424 shares of our common stock, valued at $0.1 million, in accordance with the anti-dilution provisions of the OSIF License Agreement. The OSIF License Agreement has been subsequently amended to add additional patent rights and technology disclosures to the licensed intellectual property, and we reimbursed OSIF for patent prosecution costs associated with such additional patent rights.
Pursuant to the OSIF License Agreement, we are required to pay OSIF a total of up to $4.2 million upon achievement of certain clinical development, regulatory approval, and commercial sales milestones,
for each licensed product. We are also required to pay OSIF tiered low-single digit running royalties based on annual net sales of each licensed product, along with a specified minimum annual royalty amount for each licensed product, subject to customary reductions and offsets. In addition, we are responsible for certain patent costs incurred after the effective date of the OSIF License Agreement and annual maintenance fees. The OSIF License Agreement also requires us to pay OSIF tiered percentages of certain non-royalty sublicense consideration, ranging from a low-twenties to mid-twenties percentage at earlier stages of development and a low-tens to mid-teens percentage at later stages of development, with the applicable percentage depending on the stage of development at which the sublicense is granted. In connection with the Biogen Agreement, during the year ended December 31, 2025, we paid OSIF $0.3 million in sublicensing fees related to the upfront payment received under the Biogen Agreement and no other payments were made for any other period. No milestone payments or net sales-based royalties had become due to OSIF through June 30, 2026. No repurchase rights of common stock issued to OSIF had become payable as of June 30, 2026.
Purchase and Other Obligations
We enter into contracts in the normal course of business with CROs and other third-party vendors for preclinical and commercial supply manufacturing, support for pre-commercial activities, research, and development activities, and other services and products for our operations. These contracts are generally cancelable upon written notice. Payments due upon cancellation consist generally of payments for services provided and expenses incurred up to the date of cancellation.
Lease Obligations
In December 2023, we entered into a lease of office and laboratory space in Cambridge, Massachusetts, which became our corporate headquarters. In June 2025, we entered into the First Amendment to Lease (“Amendment”), which expanded the leased premises to include the entire second floor and a portion of the fourth floor, provided for the surrender of the original first floor and penthouse premises, and extended the lease term through September 30, 2031. Following the Amendment, our leased premises consist of approximately 51,726 rentable square feet. Under the terms of the Amendment, the lease term for the retained and substitute premises expires on September 30, 2031. The lease includes a renewal option which extends the term of the lease for an additional 36 months, however, we have determined that it is not reasonably certain that we will exercise this option.
Monthly rent payments under the lease, which include base rent charges of approximately $0.5 million per month, are subject to yearly rent increases of 3% through the end of the lease term. Base rent is fully abated during the months of December 2025, December 2026, and December 2027. The Company is responsible for paying a pro rata share of costs incurred for common area maintenance, real estate taxes, and property insurance related to the leased space, which are considered variable lease payments and are expensed as incurred. The total rent payments to be paid over the non-cancelable term of this lease are $39.6 million. We have a weighted-average incremental borrowing rate of 14.0% and a weighted-average remaining lease term of 5.7 years.
Refer to Note 7 to our audited consolidated financial statements included elsewhere in this prospectus for more information on our lease obligations.
Critical Accounting Estimates and Significant Judgments
Our management’s discussion and analysis of our financial condition and results of operations is based on our consolidated financial statements, which have been prepared in accordance with GAAP. The preparation of these financial statements requires us to make estimates and judgments that affect the reported amounts of assets, liabilities, and expenses and the related disclosure of contingent assets and liabilities in our financial statements. We base our estimates on historical experience, known trends and events, and various other factors that we believe to be reasonable under the circumstances, the results of which form the basis for making judgments about the carrying values of assets and liabilities that are not readily apparent from other sources. We evaluate our estimates and judgments on an ongoing basis. Our actual results may differ from these estimates under different assumptions or conditions.
While our significant accounting policies are described in more detail in Note 2 to our audited consolidated financial statements included elsewhere in this prospectus, we believe that the following accounting policies are those most critical to the judgments and estimates used in the preparation of our consolidated financial statements.
Revenue Recognition
To date, our revenue is generated from our research, collaboration and license agreements with our collaboration partners. Under each agreement, we identify the performance obligations and determine the transaction price at contract inception, including assessment of variable consideration. We recognize revenue over time as we perform the research services using an input method which is based on the actual costs incurred relative to total costs expected to be incurred. The transaction price is updated at each reporting date for any changes in estimated variable consideration. Significant judgment is required in (i) identifying the performance obligations under each agreement, including assessing whether promises are distinct, (ii) estimating variable consideration related to cost reimbursements, milestones, and other contingent payments, including the constraint on variable consideration, and (iii) estimating the total costs expected to be incurred over the performance period. Changes in these estimates can have a material impact on the timing and amount of revenue recognized.
Research and Development
As part of the process of preparing our consolidated financial statements, we are required to estimate our accrued research and development expenses as of each balance sheet date. In accruing these costs, we estimate based on the work completed and in accordance with agreements established with our third-party service providers under the service agreements. This process involves reviewing open contracts and purchase orders, communicating with internal personnel to identify services that have been performed on our behalf and estimating the level of service performed and the associated cost incurred for the service when we have not yet been invoiced or otherwise notified of the actual cost. We recognize direct development costs based on an evaluation of the progress to completion of specific tasks using information provided to us by our vendors or our estimate of the level of service that has been performed at each reporting date. Payments for these activities are based on the terms of the individual agreements, which may differ from the timing of when costs are incurred. In circumstances where amounts have been paid in excess of costs incurred, we record a prepaid expense.
Although we do not expect our estimates to be materially different from amounts actually incurred, if our estimates of the status and timing of services performed differ from the actual status and timing of services performed, it could result in us reporting amounts that are too high or too low in any particular period. To date, there have been no material differences between our estimates of such expenses and the amounts actually incurred.
Stock-Based Compensation
We periodically grant equity-based payment awards in the form of stock options to employees, directors and non-employee consultants and record stock-based compensation expenses for awards of stock-based payments based on their estimated fair value at the grant date. For equity-based awards with service-based vesting conditions, we recognize compensation expense on a straight-line basis. For equity-based awards with performance-based vesting conditions, we recognize compensation expense at the point in time when achievement of the performance condition becomes probable. Estimating the fair value of equity awards as of the grant date using valuation models, such as the Black-Scholes option pricing model, is affected by assumptions regarding a number of variables, including the risk-free interest rate, the expected stock price volatility, the expected term of stock options, the expected dividend yield and the fair value of the underlying common stock on the date of grant. These inputs are subjective and generally require significant analysis and judgment to develop. Forfeitures are accounted for as they occur. Changes in the assumptions can materially affect the fair value and ultimately how much stock-based compensation expense is recognized. The fair value of time-vesting restricted common stock awards is equal to the common stock price on the date of grant.
We classify stock-based compensation expense in the statement of operations in the same manner in which the award recipients’ payroll costs are classified or in which the award recipients’ service payments are classified. We expect to continue to grant equity-based awards in the future, and to the extent that we do, our stock-based compensation expense recognized in future periods will likely increase.
See Note 12 to our audited consolidated financial statements included elsewhere in this prospectus for further details.
Determination of the Fair Value of Common Stock
As there is no public market for our common stock to date, the fair value of the shares of our common stock underlying these share-based awards has been determined by our board of directors, as of the date of each stock option grant, by considering a number of objective and subjective factors, including third-party valuations of our common stock, the valuation of comparable companies, our operating and financial performance, and general and industry-specific economic outlook, among other factors. The assumptions underlying these valuations represented our best estimate, with the assistance of a third-party valuation specialist, which involved inherent uncertainties and the application of our judgment. These third-party valuations were performed in accordance with the guidance outlined in the American Institute of Certified Public Accountants’ Accounting and Valuation Guide, Valuation of Privately Held Company Equity Securities Issued as Compensation. As a result, if we had used different assumptions or estimates, the fair value of our common stock and the stock-based compensation expense could have been materially different.
Our common stock valuations were prepared using either an option pricing method (“OPM”) or a hybrid method, both of which used market approaches to estimate our enterprise value. The OPM treats common stock and convertible preferred stock as call options on the total equity value of a company, with exercise prices based on the value thresholds at which the allocation among the various holders of a company’s securities changes. Under this method, the common stock has value only if the funds available for distribution to stockholders exceeded the value of the convertible preferred stock liquidation preferences at the time of the liquidity event, such as a strategic sale or a merger. The hybrid method is a probability-weighted expected return method (“PWERM”), where the equity value in one or more of the scenarios is calculated using an OPM. The PWERM is a scenario-based methodology that estimates the fair value of common stock based upon an analysis of future values for us, assuming various outcomes. The common stock value is based on the probability-weighted present value of expected future investment returns considering each of the possible outcomes available as well as the rights of each class of stock. The future value of the common stock under each outcome is discounted back to the valuation date at an appropriate risk-adjusted discount rate and probability weighted to arrive at an indication of value for the common stock. A discount for lack of marketability (“DLOM”) of the common stock is then applied to arrive at an indication of value for the common stock.
These third-party valuations were performed at various dates, which resulted in valuations of our common stock at $3.83 per share as of April 17, 2024, $1.84 per share as of April 8, 2025, $5.36 per share as of May 20, 2026, $7.89 per share as of June 5, 2026 and $9.10 per share as of July 23, 2026. Given the absence of a public trading market, our board of directors, with input from management, considered the results of these third-party valuations in addition to numerous objective and subjective factors to determine the fair value of our common stock. The factors included, but were not limited to:
•
the prices at which we sold convertible preferred stock to new and existing investors and the rights and preferences of the convertible preferred stock relative to our common stock at the time of each grant;
•
our ability to raise future financings;
•
the progress of our research and development efforts, including the status of clinical development for our product candidates;
•
our financial position, including cash on hand, and our historical and forecasted performance and operating results;
•
our stage of development and business strategy and the material risks related to our business and industry;
•
the achievement of enterprise milestones, including entering into license agreements;
•
any external market conditions affecting the biotechnology industry and trends within the biotechnology industry;
•
the likelihood of achieving a liquidity event for the holders of our convertible preferred stock and holders of our common stock, such as an initial public offering, or a sale of our Company, given prevailing market conditions; and
•
the analysis of initial public offerings and the market performance of similar companies in the biopharmaceutical industry.
The assumptions underlying these valuations were highly complex and subjective and represented management’s best estimates, which involved inherent uncertainties and the application of management’s judgment. As a result, if we had used significantly different assumptions or estimates, the fair value of our common stock and our stock-based compensation expense could have been materially different. Once a public trading market for our common stock has been established in connection with the completion of this offering, it will no longer be necessary for our board of directors to estimate the fair value of our common stock in connection with our accounting for granted stock options and other equity awards we may grant, as the fair value of our common stock will be determined based on the quoted market price of our common stock.
The following table summarizes by grant date the number of shares subject to awards granted under our 2023 Plan from January 1, 2025 through the date of this prospectus, the per share exercise price of the awards and weighted average grant date fair value on each grant date:
|
Grant Date
|
|
|
Type of Award
|
|
|
Number of
Shares
Subject
to Award(1)
|
|
|
Per Share
Exercise Price
of Award
|
|
|
Per Share
Fair Value
of Common
Stock on
Grant Date
|
|
|
Weighted Average
Grant Date Fair
Value Per Share
|
|
|
February 25, 2025
|
|
|
Stock Option
|
|
|
|
|
373,437 |
|
|
|
|
$ |
0.92 |
|
|
|
|
$ |
3.83(2) |
|
|
|
|
$ |
3.47 |
|
|
|
May 13, 2025
|
|
|
Stock Option
|
|
|
|
|
834,592 |
|
|
|
|
$ |
1.84 |
|
|
|
|
$ |
1.84 |
|
|
|
|
$ |
1.43 |
|
|
|
August 11, 2025
|
|
|
Stock Option
|
|
|
|
|
314,654 |
|
|
|
|
$ |
1.84 |
|
|
|
|
$ |
1.84 |
|
|
|
|
$ |
1.42 |
|
|
|
November 21, 2025
|
|
|
Stock Option
|
|
|
|
|
71,388 |
|
|
|
|
$ |
1.84 |
|
|
|
|
$ |
1.84 |
|
|
|
|
$ |
1.43 |
|
|
|
February 13, 2026
|
|
|
Stock Option
|
|
|
|
|
1,681,287 |
|
|
|
|
$ |
1.84 |
|
|
|
|
$ |
1.84 |
|
|
|
|
$ |
1.45 |
|
|
|
February 25, 2026
|
|
|
Stock Option
|
|
|
|
|
53,810 |
|
|
|
|
$ |
1.84 |
|
|
|
|
$ |
1.84 |
|
|
|
|
$ |
1.46 |
|
|
|
May 12, 2026
|
|
|
Stock Option
|
|
|
|
|
163,342 |
|
|
|
|
$ |
5.36 |
|
|
|
|
$ |
5.36 |
|
|
|
|
$ |
4.37 |
|
|
|
June 4, 2026
|
|
|
Stock Option
|
|
|
|
|
630,347 |
|
|
|
|
$ |
5.36 |
|
|
|
|
$ |
7.89 (3) |
|
|
|
|
$ |
6.79 |
|
|
|
July 8, 2026
|
|
|
Stock Option
|
|
|
|
|
375,616 |
|
|
|
|
$ |
7.89 |
|
|
|
|
$ |
7.89 |
|
|
|
|
$ |
5.85 |
|
|
|
July 8, 2026
|
|
|
Restricted
Common Stock
|
|
|
|
|
109,828 |
|
|
|
|
$ |
7.89 |
|
|
|
|
$ |
7.89 |
|
|
|
|
$ |
7.89 |
|
|
|
August 4, 2026
|
|
|
Stock Option
|
|
|
|
|
612,842 |
|
|
|
|
$ |
9.10 |
|
|
|
|
$ |
9.10 |
|
|
|
|
$ |
6.78 |
|
|
(1)
Each grant date includes both time-based and performance-based stock options granted at the exercise price shown. For the year ended December 31, 2025, the 373,437 options granted at a $0.92 exercise price consisted of 232,033 time-based and 141,404 performance-based options, and the 1,220,634 options granted at a $1.84 exercise price consisted of 898,433 time-based and 322,201 performance-based options. For the six months ended June 30, 2026, the 1,735,097 options granted at a $1.84 exercise price consisted of 512,931 time-based and 1,222,166 performance-based options, the 163,342 options granted at a $5.36 exercise price consisted of 129,408 time-based and 33,934 performance-based options, the 630,347 options granted at a $5.36 exercise price consisted of 621,123 time-based and 9,224 performance-based options. For the period after June 30, 2026, the 375,616 options granted at a $7.89 exercise price consisted of 151,564 time-based and 224,052 performance-based options, the 109,828 restricted common stock granted at a $7.89 exercise price consisted of 27,456 time-based and 82,372 performance-based restricted common stock, and the 612,842 options granted at a $9.10 exercise price consisted of 188,466 time-based and 424,376 performance-based options.
(2)
At the time of the stock option grant on February 25, 2025, our board of directors determined that the fair value of our common stock of $0.92 per share reasonably reflected the fair value of our common stock as of the grant date. However, the fair value of our common stock as of the date of this grant was adjusted in connection with a retrospective fair value assessment for accounting purposes.
(3)
At the time of the stock option grant on June 4, 2026, our board of directors determined that the fair value of our common stock of $5.36 per share reasonably reflected the fair value of our common stock as of the grant date. However, the fair value of our common stock as of the date of this grant was adjusted in connection with a retrospective fair value assessment for accounting purposes.
The fair value of our common stock of $3.83 per share as of February 25, 2025 was determined by our board of directors, based, in part, on the $3.83 per share value indicated in the retrospective third-party valuation prepared as of April 17, 2024. In particular, the valuation determined our enterprise value using an OPM backsolve approach that was primarily based on the $10.0295 price per share paid by new and existing investors in the closing of our Series A convertible preferred stock in April 2024. A DLOM of the common stock was then applied to arrive at an indication of value for our common stock.
The fair value of our common stock of $1.84 per share from May 13, 2025 to February 25, 2026 was determined by our board of directors, based, in part, on the $1.84 per share value indicated in the third-party valuation prepared as of April 8, 2025. In particular, the valuation determined our enterprise value using an OPM market-adjusted backsolve approach that was based on the $10.0295 price per share paid by new and existing investors in the closing of our Series A preferred stock in April 2024. The market adjustment applied to the equity value considered the performance of guideline public companies and the biotech indices since the most recent sale of our convertible preferred stock through the valuation date. A DLOM of the common stock was then applied to arrive at an indication of value for our common stock.
The fair value of our common stock of $5.36 per share from May 12, 2026 to June 3, 2026 was determined by our board of directors, based, in part, on the $5.36 per share value indicated in the third-party valuation prepared as of May 20, 2026. In particular, the valuation determined our enterprise value using an OPM backsolve approach that was based on the $14.63 price per share paid by new and existing investors in the closing of our Series B convertible preferred stock in May and June 2026. A DLOM of the common stock was then applied to arrive at an indication of value for our common stock.
The fair value of our common stock of $7.89 per share from June 4, 2026 to July 7, 2026 was determined by our board of directors, and by us, based, in part, on the $7.89 per share value indicated in the third-party valuation prepared as of June 5, 2026 and our decision to apply the June 5, 2026 valuation to our June 4, 2026 grant for accounting purposes. In particular, the valuation determined our enterprise value using the hybrid method, which included a PWERM, with an initial public offering (“IPO”) scenario, and a sale scenario. Our enterprise value in the IPO scenario was based on guideline IPO transactions identified within the last two years, which was adjusted by a risk adjusted discount rate. The IPO scenario also assumed an estimated timeline for the IPO to occur. Our enterprise value for the sale scenario was based on an OPM backsolve method based on the $14.63 price per share paid by new and existing investors in the closing of our Series B preferred stock in May 2026 and June 2026. A DLOM of the common stock was then applied to arrive at an indication of value for our common stock.
The fair value of our common stock of $9.10 per share as of July 23, 2026 was determined by our board of directors, based, in part, on the $9.10 per share value indicated in the third-party valuation prepared as of July 23, 2026. In particular, the valuation determined our enterprise value using the hybrid method, which included a PWERM, with an IPO scenario, and a sale scenario. Our enterprise value in the IPO scenario was based on guideline IPO transactions identified within the last two years, which was adjusted by a risk adjusted discount rate. The IPO scenario also assumed an estimated timeline for an IPO to occur. Our enterprise value for the sale scenario was based on an OPM market-adjusted backsolve method based on the $14.63 price per share paid by new and existing investors in the closing of our Series B convertible preferred stock in May 2026 and June 2026. The market adjustment applied to the equity value considered the performance of guideline public companies and the biotech indices since the most recent sale of our preferred stock through the valuation date. A DLOM of the common stock was
then applied to arrive at an indication of value for our common stock. In addition, our board of directors determined that the fair value of our common stock remained at $9.10 per share through August 4, 2026.
Off-Balance Sheet Arrangements
During the periods presented, we did not have, nor do we currently have, any off-balance sheet arrangements as defined in the rules and regulations of the SEC.
Recent Accounting Pronouncements
See Note 2 to our audited consolidated financial statements included elsewhere in this prospectus for a description of recent accounting pronouncements applicable to our consolidated financial statements.
Emerging Growth Company and Smaller Reporting Company Status
We are an “emerging growth company,” as defined in Section 2(a) of the Securities Act, as modified by the JOBS Act. As such, we are eligible to take advantage of certain exemptions from various reporting requirements that are applicable to other public companies that are not emerging growth companies including:
•
being permitted to provide only two years of audited financial statements in addition to any required unaudited interim financial statements and a correspondingly reduced “Management’s Discussion and Analysis of Financial Condition and Results of Operations” disclosure in this prospectus;
•
not being required to comply with the auditor attestation requirements in the assessment of our internal control over financial reporting pursuant to Section 404 of the Sarbanes-Oxley Act;
•
reduced disclosure obligations regarding executive compensation;
•
exemptions from the requirements of holding a nonbinding advisory vote on executive compensation and shareholder approval of any golden parachute payments not previously approved; and
•
exemptions from compliance with the requirements of the Public Company Accounting Oversight Board regarding the communication of critical audit matters in the auditor’s report on the financial statements.
Further, Section 102(b)(1) of the JOBS Act exempts emerging growth companies from being required to comply with new or revised financial accounting standards until private companies (that is, those that have not had a Securities Act registration statement declared effective or do not have a class of securities registered under the Exchange Act) are required to comply with the new or revised financial accounting standards. The JOBS Act provides that an emerging growth company can elect to opt out of the extended transition period and comply with the requirements that apply to non-emerging growth companies but any such election to opt out is irrevocable. We have elected not to opt out of such extended transition period which means that when a standard is issued or revised and it has different application dates for public or private companies, we, as an emerging growth company, will adopt the new or revised standard whenever such adoption is required for private companies. We may also choose to early adopt any new or revised accounting standards whenever such early adoption is permitted. This may make comparison of our financial statements with another public company, which is neither an emerging growth company nor an emerging growth company that has opted out of using the extended transition period, difficult or impossible because of the potential differences in accounting standards used.
We will remain an emerging growth company until the earliest of (i) the last day of the fiscal year following the fifth anniversary of the completion of this initial public offering, (ii) the last day of the fiscal year in which we have total annual gross revenue of at least $1.235 billion, (iii) the last day of the fiscal year in which we are deemed to be a “large accelerated filer” as defined in Rule 12b-2 under the Exchange Act, which would occur if the market value of the common stock held by non-affiliates exceeded
$700.0 million as of the last business day of the second fiscal quarter of such year, or (iv) the date on which we have issued more than $1.0 billion in non-convertible debt securities during the prior three‑year period.
We are also a “smaller reporting company” as defined in the Exchange Act. We may continue to be a smaller reporting company even after we are no longer an emerging growth company. We may take advantage of certain of the scaled disclosures available to smaller reporting companies and will be able to take advantage of these scaled disclosures for so long as (i) the market value of our voting and non-voting common stock held by non-affiliates is less than $250.0 million measured on the last business day of our second fiscal quarter, or (ii)(a) our annual revenue is less than $100.0 million during the most recently completed fiscal year, and (b) the market value of our voting and non-voting common stock held by non-affiliates is less than $700.0 million measured on the last business day of our second fiscal quarter.
BUSINESS
Overview
We are a clinical-stage biotechnology company developing the next generation of RNA interference (“RNAi”) therapeutics designed to silence the expression of disease-relevant proteins, with the aim of delivering meaningful, durable benefits for patients in need. We believe our deep RNAi expertise and City Platform capabilities uniquely position us to achieve our goal of developing potentially best-in-class therapeutics. Our vision is to lead the next generation of RNAi therapeutics, and our mission is to improve patients’ lives through breakthrough RNAi science as we build a biopharmaceutical company with enduring impact.
Our current pipeline is focused on indications with significant unmet medical need, including thromboembolic diseases, Stargardt disease type 1 (“Stargardt disease”), and anemia of chronic disease, where we believe disease biology supports RNAi-mediated knockdown as a differentiated approach compared to other modalities. Led by pioneers and experts in the field of RNAi, we have made rapid progress since founding our company in July 2023, discovering and advancing three programs into or toward the clinic in under three years, and our goal is for our City Platform to deliver one to two new investigational new drug application (“IND”) or IND-equivalent applications per year.
Several members of our leadership team, including John Maraganore, Ph.D., our co-founder and Executive Chair and former founding Chief Executive Officer of Alnylam Pharmaceuticals, Inc. (“Alnylam”), and Tracy Zimmermann, Ph.D., our Chief Scientific Officer, played a foundational role in establishing RNAi therapeutics as a recognized treatment modality, as evidenced by multiple FDA-approved RNAi therapeutics that each of them worked in development on, and our President and Chief Executive Officer, Andrew (Andy) Orth, M.B.A., led the launch of multiple RNAi therapeutics, including the first approved RNAi therapy. The team at City has contributed to the discovery, development, or commercialization of seven of the eight currently U.S. Food and Drug Administration (“FDA”)-approved RNAi therapeutics, and 75% of our research leaders have prior RNAi therapeutics discovery or development experience.
RNAi is a naturally occurring biological process that regulates or silences gene expression by “interfering” with messenger RNA (“mRNA”) before it can be translated into protein. RNAi has inherent advantages as a therapeutic mechanism because it is a highly specific, potent, and durable approach to selectively silence the expression of disease-relevant proteins, including intracellular targets that are inaccessible to traditional small molecules and antibodies. In addition, RNAi therapeutics benefit from a simpler molecular architecture relative to biologics, consisting of short, chemically synthesized oligonucleotides rather than large, complex proteins, which enables more consistent and scalable manufacturing. RNAi has been established mechanistically and commercially, with the first of the eight approved RNAi therapeutics coming to market in 2018 and multiple products achieving substantial and growing commercial sales. With approximately 100 programs currently in clinical development, we believe RNAi therapeutics are poised to follow a growth trajectory analogous to that of monoclonal antibody therapeutics, a modality that was first approved by the FDA in the mid-1990s and has since expanded to more than 150 approved medicines today.
Currently approved RNAi therapeutics have focused on diseases that can be treated through delivery to liver cells. RNAi therapeutics typically consist of a short, chemically synthesized RNAi trigger molecule that initiates the RNAi process to silence a target gene. Current RNAi technologies predominantly utilize a N-acetylgalactosamine (“GalNAc”) ligand to deliver this therapeutic trigger to liver cells, and this approach is well-established in preclinical and clinical studies, and in subsequent regulatory approvals. However, while the first generation of RNAi therapeutics has been successful, significant opportunities remain to further improve and extend the reach of RNAi therapeutics. To unlock the true potential of RNAi therapeutics, new technologies are needed to extend delivery beyond the liver to the more than 200 distinct cell types and thousands of diseases that remain beyond the reach of current RNAi approaches.
We are aiming to define the future of the field of RNAi therapeutics both in and beyond the liver. We believe the next major advances in RNAi will come from innovations in RNAi trigger molecule design, tissue-selective delivery, and target selection. We have developed our City Platform to integrate
these capabilities, engineering our RNAi therapeutic candidates for liver-directed applications while also extending the reach of the modality to extrahepatic tissues. Together with our deep RNAi expertise and disciplined execution, our City Platform positions us to develop RNAi therapeutics that have the potential to make a meaningful and durable difference for patients while supporting the advancement of our goal of filing one to two new IND or IND-equivalent applications per year. We are currently advancing our three lead product candidates for the treatment of diseases where there is significant unmet medical need.
CITY-FXI
Our most advanced product candidate, CITY-FXI, is a City-engineered GalNAc-conjugated small interfering RNA (“siRNA”) therapeutic candidate targeting Factor XI (“FXI”). CITY-FXI illustrates our strategy of engineering highly potent and specific RNAi triggers directed at therapeutic targets supported by third-party human genetic evidence and clinical data. We are advancing CITY-FXI for the prevention of thromboembolic diseases based on its potential to reduce FXI activity and thereby prevent thromboembolic events while mitigating bleeding risk. However, CITY-FXI is in early clinical development, and we have not demonstrated that CITY-FXI is safe or effective, provides advantages over existing or investigational therapies or will receive regulatory approval. Thrombosis, the formation of harmful clots inside a blood vessel, is responsible for approximately one in four deaths worldwide. It is estimated that 40 to 50 million patients are at risk for thromboembolic diseases across indications, including atrial fibrillation (“AFib”), prior venous thromboembolism (“VTE”), peripheral artery disease, and coronary artery disease. The global anticoagulants market was estimated at approximately $37 billion in 2025 and is projected to reach approximately $56 billion by 2033, with FXI-targeted therapeutics representing the fastest-growing segment. Many patients remain untreated or undertreated due to bleeding concerns, contraindications, treatment burden and adherence challenges, underscoring the need for new treatment options despite the availability of direct oral anticoagulants (“DOACs”), which is the current standard of care for many thromboembolic diseases.
FXI is a therapeutic target supported by human genetic and clinical data demonstrating that it contributes to pathological thrombosis while playing a limited role in hemostasis, or normal blood clotting. CITY-FXI aims to prevent the expression of the protein itself, enabling a potentially complete and specific reduction of FXI function with a convenient biannual subcutaneous dosing profile. In preclinical studies with CITY-FXI, we observed 95% FXI knockdown with prolonged clotting time in non-human primates (“NHPs”) and no observed off-target effects in cell-based assays. We initiated our Phase 1 clinical trial for CITY-FXI in 2026. Initial data from the first three cohorts of the single-ascending dose (“SAD”) portion of our study in healthy volunteers showed that CITY-FXI was generally well-tolerated by participants with greater than 60% FXI knockdown observed following a single 15 mg subcutaneous dose, close to 80% FXI knockdown after a single 50 mg subcutaneous dose, and 85% knockdown three weeks after a single 150 mg subcutaneous dose. We expect to report additional data from the SAD portion of our Phase 1 study in late 2026. We intend to develop CITY-FXI for thromboembolic disease indications, including secondary stroke prevention, DOAC-ineligible AFib, extended secondary prophylaxis following VTE, and cancer-associated thrombosis, specifically in patients for whom DOACs are often not used because of heightened bleeding risk. We plan to first demonstrate clinical efficacy and safety in thrombotic disease through a Phase 2 trial in total knee arthroplasty (“TKA”), which we expect to initiate in mid-2027.
CITY-RBP4
Our second product candidate, CITY-RBP4, is a City-engineered GalNAc-conjugated siRNA therapeutic in development for the treatment of Stargardt disease, which is the most common inherited macular degenerative disease in children and young adults. There are currently no approved therapies for the disease. More than 30,000 people in the United States are diagnosed with Stargardt disease, and some studies suggest that the prevalence is even higher, indicating there could be a significant number of undiagnosed patients who emerge upon FDA approval of a treatment. Stargardt disease is debilitating, and many patients become legally blind in their late twenties. Patients face decades of progressive loss of central vision that renders daily activities, including reading, recognizing faces, and driving impossible.
Stargardt disease is caused by mutations in the ABCA4 gene, which lead to the toxic accumulation of vitamin A byproducts in the eye. Retinol binding protein 4 (“RBP4”) serves as the primary carrier of retinol, a form of vitamin A, that transports retinol from the liver to the eye. RBP4 is supported by clinical proof-of-mechanism data as an approach for the treatment of Stargardt disease. CITY-RBP4 targets RBP4 in the liver to reduce the toxic accumulation of vitamin A byproducts in the eye associated with disease progression.
In a preclinical mouse model of Stargardt disease, suppression of RBP4 with an siRNA targeting mouse RBP4 reduced A2E, a major toxic vitamin A byproduct, to normal levels at therapeutically relevant doses. Further, in a preclinical study in NHPs, following treatment with CITY-RBP4, we observed approximately 90% suppression of circulating vitamin A and RBP4 at dose levels of 3 mg/kg and 10 mg/kg. Additionally, CITY-RBP4’s long half-life, as supported by pharmacodynamic modeling data, suggests the potential for quarterly to biannual subcutaneous dosing in humans. We filed an application in mid-2026 and plan to initiate a Phase 1 clinical trial for CITY-RBP4 in Stargardt disease in late 2026. We expect to report initial data from the Phase 1 clinical trial in mid-2027. We are also exploring CITY-RBP4 in adjacent indications, including Geographic Atrophy (“GA”) secondary to age-related macular degeneration. We expect to initiate a Phase 2 clinical trial for CITY-RBP4 in GA in late 2027.
CITY-TFR2
Our third program, CITY-TFR2, is a City-engineered GalNAc-conjugated siRNA therapeutic in development for anemia of chronic disease, which accounts for up to 40% of all anemias and affects more than five million people in the United States. Current therapies, including erythropoiesis-stimulating agents (“ESAs”) and hypoxia-inducible factor prolyl hydroxylase inhibitors (“HIF-PH inhibitors”), come with significant safety and efficacy limitations, and many patients become dependent on blood transfusions. Patients with anemia of chronic disease often experience debilitating fatigue, weakness, and reduced physical function that can significantly diminish their quality of life.
In inflammatory states associated with anemia of chronic disease, hepcidin, the hormone that controls systemic iron, becomes elevated, restricting iron availability and starving red blood cell production, thereby causing anemia. CITY-TFR2 targets transferrin receptor 2 (“TFR2”), a protein predominantly expressed in the liver that is critical in signaling pathways governing hepcidin production and is a compelling therapeutic target for anemia of chronic disease, supported by human genetic data. By reducing TFR2 expression, CITY-TFR2 is designed to lower hepcidin levels, restore iron availability, and address a central driver of anemia in patients with anemia of chronic disease.
In a preclinical study in NHPs, following treatment with CITY-TFR2, we observed approximately 90% TFR2 mRNA knockdown that led to an increase in serum iron and transferrin saturation, with durations of action that suggest the potential for quarterly to biannual subcutaneous dosing in humans. Our current plan is to initially focus CITY-TFR2 on anemia associated with myelofibrosis, as anemia is a major contributor to the clinical burden of the disease. In July 2026, we entered into an investigator-initiated trial (“IIT”) study agreement with Overland Therapeutics (SH) Co. Ltd. (“Overland”), a biotechnology company that has operations in the United States and China, to support the conduct of an IIT for CITY-TFR2 in myelofibrosis-associated anemia. The IIT is expected to begin in China in late 2026, with a goal of demonstrating clinical proof-of-concept. Pursuant to the agreement, we will provide support for the IIT, supply CITY-TFR2 and fund covered study-related services and expenses, and Overland, as local study sponsor, will provide clinical and operational support in connection with the IIT, which will be conducted by investigators at participating medical institutions in China. While the IIT is not one of our traditional company-sponsored clinical trials, subject to applicable law and the applicable agreements with the investigators and medical institutions, data and results generated from the IIT will be provided to us. Although data from the IIT may support our development program for CITY-TFR2, including by providing initial clinical proof-of-concept data and informing our planned Phase 1/2 clinical trial, there can be no assurance that the FDA or comparable foreign regulatory authorities will accept such data in support of future regulatory submissions or marketing approval. We expect to report data from the IIT in mid-2027. We plan to file an IND or IND-equivalent application in late 2026 and initiate a Phase 1/2 study for CITY-TFR2 in early 2027. Beyond this initial indication, we see potential
to expand into other chronic conditions in which hepcidin levels drive anemia with severe adverse outcomes, such as chronic kidney disease and chronic inflammatory disorders, including inflammatory bowel disease.
Partnerships
We have formed strategic collaborations with Bausch + Lomb Ireland Limited (“Bausch + Lomb”) and Biogen MA Inc. (“Biogen”) to advance extrahepatic RNAi therapeutics for diseases of the eye and the central nervous system (“CNS”), respectively. We believe these partnerships reinforce the potential of and enhance our City Platform and expertise, as well as the potential of our next-generation RNAi trigger and delivery technologies. These arrangements also preserve our ability to pursue additional targets in the eye and CNS and retain the rights to any City Platform innovations developed from our work under the collaborations.
In July 2024, we established a strategic collaboration with Bausch + Lomb, which is focused on the development of a novel therapy for the treatment of retinal diseases, including GA. The collaboration leverages our next-generation ligand delivery technologies to enable intravitreal administration for the development of a novel RNAi therapy directed against a specific disease target in the retina. We plan to substantially complete the research and development under the B+L Agreement to enable Bausch + Lomb to select a development candidate by late 2026.
In May 2025, we established a strategic collaboration with Biogen that combines our RNAi trigger technology with Biogen’s proprietary drug delivery technology to develop a novel systemic therapy for the treatment of certain CNS diseases. The collaboration initially focuses on a single target, with an option to designate one additional target for a separate collaboration and license agreement on substantially the same terms. We plan to substantially complete the research and development under the Biogen Agreement by late 2026 to enable Biogen to select a development candidate by late 2027.
Discovery Candidates
Beyond our lead programs, we are progressing discovery candidates designed to expand the reach of RNAi therapeutics into new tissues, targets, and disease areas. We have a fourth liver-directed program, CITY-L04, with an undisclosed target as well as two discovery-stage programs based on our proprietary cleavage-inducing tiny RNAs (“cityRNAs”), which comprise CITY-O02 and CITY-C02 and are focused on the treatment of ocular and CNS indications, respectively. Our cityRNA triggers are approximately 30% shorter and carry less negative charge than traditional siRNAs. These properties offer several advantages including improved tissue penetration and cellular uptake that may increase potency, specificity and durability, while enabling delivery beyond the liver to other tissue types.
The following table summarizes our current development stage programs:
Platform
The City Platform is the engine behind every therapeutic we develop. Our City Platform is designed to integrate three core capabilities: RNAi trigger molecule engineering, novel delivery conjugates that allow for tissue selectivity, and target discovery and selection enabled by our insights into human genetics. Critical to every RNAi medicine is the trigger that harnesses the cellular RNAi machinery to silence a disease-relevant gene. We believe our expertise enables us to engineer the trigger molecule for high potency and specificity, and our novel cityRNA triggers are a differentiated approach to unlock extrahepatic delivery. Our proprietary artificial intelligence and machine learning (“AI/ML”)-enabled trigger design capability uses a large RNAi activity dataset to predict high-potential trigger molecules in silico, accelerating design cycles from months to weeks. We conjugate our triggers with tissue-targeting ligands for precise delivery to cell types in and beyond the liver, starting with our proprietary LRP1 ligands, which are designed to enable delivery to ocular, CNS, and adipose tissues. Our pipeline programs are selected through a genetics-led target discovery and selection capability that leverages insights from human biology and other sources of substantiation, including clinical data. We use this capability to identify differentiated genetic targets where RNAi may be able to confer a durable therapeutic advantage. By uniting these capabilities in a single platform, we aim to systematically create and develop potentially best-in-class therapeutics that support our goal of filing one to two new IND or IND-equivalent applications per year. The filing of an IND or IND-equivalent application, or the clearance of any such application by the FDA or a comparable foreign regulatory authority, is not indicative of the future success of any of our product candidates and does not increase the likelihood that any of our product candidates will receive regulatory approval. Given our early stage of development, it will take many years before we complete clinical development of and receive regulatory approval for any of our product candidates, if ever.
Certain foundational elements of our City Platform are supported by intellectual property licensed from the Ohio State Innovation Foundation (“OSIF”). In October 2023, we entered into a license agreement (as amended, the “OSIF License Agreement”) with OSIF, pursuant to which OSIF granted us an exclusive, royalty-bearing license to certain patent rights covering the originating technology underlying cityRNAs and certain other aspects of our technology platform.
Our Team and History
Defining the next generation of RNAi therapeutics requires a unique combination of scientific, clinical, and commercial expertise. We have assembled a team with deep experience across each of these disciplines. Collectively, our team at City has contributed to the discovery, development, or commercialization of seven of the eight currently FDA-approved RNAi therapeutics. The team at City has a proven track record of translating innovative science into transformative therapies. Members of our leadership team include:
•
John Maraganore, Ph.D., our Executive Chair, is one of our co-founders and is a pioneer in the field of RNAi therapeutics. Dr. Maraganore was the founding Chief Executive Officer and a Director of Alnylam, which he led for nearly 20 years. At Alnylam, Dr. Maraganore built and led the company from early platform research through global approval and commercialization of the first five RNAi therapeutics, including ONPATTRO, GIVLAARI, OXLUMO, LEQVIO, and AMVUTTRA. At Alnylam, he led the company’s value creation strategy, raising over $7.5 billion in capital, forming over 20 major pharmaceutical alliances, and creating over $25 billion in market capitalization and one of the biotechnology industry’s top companies. Prior to Alnylam, Dr. Maraganore held senior leadership roles at Millennium Pharmaceuticals and at Biogen, where he invented and led the discovery and development of ANGIOMAX (bivalirudin).
•
Andrew (Andy) Orth, M.B.A., our President and Chief Executive Officer, is a seasoned life sciences executive with more than 25 years of experience developing and commercializing novel medicines. Mr. Orth held multiple leadership roles at Alnylam, including Senior Vice President, U.S. Region, where he led Alnylam’s U.S. business and launched ONPATTRO, the first-ever RNAi therapeutic, followed by the launches of GIVLAARI and OXLUMO. Most recently, Mr. Orth served as Chief Commercial Officer of Krystal Biotech, Inc., where he led corporate strategy and growth, and oversaw the global commercialization and launch of VYJUVEK.
•
Keith Regnante, M.B.A., our Chief Financial Officer, is a seasoned finance executive with more than 25 years of experience in the biopharmaceutical industry. Prior to joining us, Mr. Regnante served as Chief Financial Officer of Keros Therapeutics, Inc., where he led the finance organization through a period of significant growth and multiple strategic financings and transactions, including the company’s initial public offering, two follow-on offerings and a global licensing agreement with Takeda Pharmaceutical Company Limited. Previously, Mr. Regnante served as Chief Financial Officer of Wave Life Sciences Ltd. Earlier in his career, he held finance leadership roles of increasing responsibility at Shire plc and Biogen Inc., spanning corporate finance, global research and development finance, investor relations and business development finance. At Biogen Inc., he contributed to several significant strategic transactions.
•
Baisong Mei, M.D., Ph.D., our Chief Medical Officer, is an experienced drug developer with over 25 years of industry experience and a strong track record of bringing innovative medicines through clinical development to regulatory approval. Dr. Mei played a critical leadership role in the development and regulatory approval of ALPROLIX, ELOCTATE, ALTUVIIIO, and the siRNA medicine QFITLIA. Dr. Mei is also a co-inventor of JIVI and ALTUVIIIO. Dr. Mei was previously Executive Vice President and Chief Medical Officer at Editas Medicine, Inc., Senior Global Project Head in rare disease and rare blood disorders at Sanofi S.A. (where he led Sanofi’s collaboration with Alnylam on siRNA drug clinical development) and Therapeutic Area Head for Hematology Clinical Development at Biogen.
•
Tracy Zimmermann, Ph.D., our Chief Scientific Officer, has more than 25 years of experience in drug development and has deep expertise in developing nucleic acid-based therapeutic platforms, including RNAi-based therapies. Dr. Zimmermann spent 15 years at Alnylam, culminating in her role as Senior Director, Research, where she oversaw numerous research roles spanning RNAi platform enablement to early program development. Dr. Zimmermann was the lead author of a landmark 2006 study, published in Nature, demonstrating the first RNAi silencing in NHPs, which was cited over 1,400 times. Dr. Zimmermann was critical to the discovery of AMVUTTRA, including as a co-inventor, and served as its Program Lead through early clinical development. Dr. Zimmermann further contributed to the advancement of ONPATTRO and GIVLAARI. Dr. Zimmermann previously served as Chief Development Officer at Generation Bio Co., where she led the pharmacology, toxicology, and DMPK groups to advance the company’s non-viral gene therapy platform from research into preclinical development for liver and extrahepatic indications.
•
Sebastian Trousil, Ph.D., our Chief Operating Officer, is one of our co-founders and a life sciences executive with extensive experience leading corporate operations, business
development, and finance. Dr. Trousil previously worked at RTW Investments LP and Flagship Pioneering, where he led multiple company creation efforts at both firms. During his time at RTW, Dr. Trousil served as Chief Operating Officer at Yarrow Biotechnology, Inc. Dr. Trousil is a co-founder of Harbinger Health, Inc., a liquid biopsy company for early cancer detection, which he led from initiation to early proof-of-concept.
From our inception in July 2023 through June 30, 2026, we have raised approximately $238.8 million in equity capital from a syndicate of premier life sciences investors, including ARCH Venture Partners, Fidelity Management & Research Company, Invus and Viking Global Investors. Prospective investors should not rely on the investment decisions of our existing investors, as these investors may have different risk tolerances and, in certain cases, have received their shares in private financings at prices lower than the price offered to the public in this offering. See the sections titled “Certain Relationships and Related Person Transactions” and “Principal Stockholders” for more information on prior purchases by and current holdings of certain of these investors.
Our Strategy
We are pioneering the next generation of RNAi therapeutics, leveraging our deep scientific expertise and proprietary platform with the goal of discovering and developing potentially best-in-class therapeutics. Our goal is to build a sustainable, fully integrated RNAi company capable of delivering transformative therapies across a broad range of diseases. As key elements of our strategy, we intend to:
•
Pioneer the future of RNAi trigger design and delivery. We intend to lead the next wave of RNAi innovation by continually advancing how RNAi therapeutics are designed and delivered. Through innovations in trigger molecule design and delivery, including with our proprietary cityRNA triggers, we are pioneering new approaches that aim to improve potency and specificity. This will enable our reach of targets beyond the liver and expand the range of tissues and diseases addressable by RNAi therapeutics. Our City Platform is designed to integrate these capabilities with the aim of overcoming longstanding challenges in the field and further expand the reach and impact of RNAi therapeutics both in and beyond the liver.
•
Advance development candidates in high unmet medical need indications where RNAi offers meaningful advantages. We are committed to progressing our pipeline across a range of indications of high unmet medical need where RNAi therapeutics have the potential to deliver meaningful benefit. By targeting diseases including thromboembolic diseases, Stargardt disease, and anemia of chronic diseases, we aim to maximize the therapeutic impact and commercial potential of our programs by addressing meaningful gaps in patient care. Our lead product candidate, CITY-FXI, is currently in a Phase 1 clinical trial, and we expect to report additional data from the SAD portion of the trial in late 2026. In addition, we filed an application for CITY-RBP4 in mid-2026 to initiate a Phase 1 clinical study in late 2026 to evaluate its safety, tolerability, pharmacokinetics, and pharmacodynamics in healthy volunteers and Stargardt patients. We also are continuing to advance CITY-TFR2 towards the clinic with an IIT expected to begin in China in late 2026, and a Phase 1/2 clinical study expected to begin in early 2027, in each case for the treatment of anemia associated with myelofibrosis.
•
Selectively pursue value-accretive partnerships to maximize the value of our portfolio. While our primary focus is the rapid and successful advancement and commercialization of our wholly owned pipeline, we also plan to continue selectively engaging in strategic partnerships that enable us to maximize the value and reach of our portfolio. Our existing collaborations with Bausch + Lomb and Biogen exemplify this approach. We plan to continue to evaluate additional collaboration, licensing, and partnership opportunities that allow us to accelerate development, access complementary capabilities, and expand our reach. At the same time, we intend to retain development and commercial rights to programs we believe we can successfully commercialize on our own, if approved.
•
Bring exceptional depth of experience to accelerate and expand our pipeline. Collectively, our team at City has contributed to the discovery, development, or commercialization of seven of the eight currently FDA-approved RNAi therapeutics, and 75% of our research leaders
have prior RNAi therapeutics discovery or development experience. Since founding our company in July 2023, we have advanced three programs into the clinic or toward planned clinical entry in late 2026. As a team of RNAi pioneers, we intend to advance the next generation of RNAi therapeutics. We combine deep expertise with the ambition to define the future of the field, and we believe our collective experience makes us uniquely positioned to navigate the scientific, regulatory, and commercial complexities of developing RNAi therapeutics efficiently and effectively. We believe our expertise and disciplined execution, coupled with our platform technology, will enable us to deliver one to two new IND or IND-equivalent application filings per year.
Our Pipeline and Programs
We are advancing a pipeline of RNAi therapeutics designed to achieve potent, specific, and durable suppression of disease-relevant hepatic and extrahepatic targets to address diseases with significant unmet medical needs. All of our product candidates are in early clinical or preclinical development. We have not demonstrated that any of our product candidates are safe or effective or provide advantages over existing or investigational therapies, and results of preclinical studies, including our NHP studies, may not be predictive of results in clinical trials. None of our product candidates has been approved by the FDA or any comparable foreign regulatory authority, and we may never obtain regulatory approval for any of our product candidates. Our three lead programs are City-engineered liver-directed siRNA therapeutic candidates in development for thromboembolic diseases, Stargardt disease, and anemia of chronic disease, where we believe disease biology supports RNAi-mediated knockdown as a differentiated approach compared to other modalities.
CITY-FXI
We intend to develop CITY-FXI for thromboembolic disease indications, including secondary stroke prevention, DOAC-ineligible AFib, extended secondary prophylaxis following VTE, and cancer-associated thrombosis, specifically in patients for whom DOACs are often not used because of heightened bleeding risk. The current standard of care for these diseases is anticoagulants, which represents a global market of approximately $37 billion in 2025 and is projected to reach approximately $56 billion by 2033. CITY-FXI is a City-engineered GalNAc-conjugated siRNA therapeutic candidate designed to suppress FXI — a therapeutic target supported by human genetic and clinical evidence demonstrating that it contributes to pathological thrombosis, or harmful blood clots, while playing a limited role in normal hemostasis. Initial data from the first three cohorts of the SAD portion of our ongoing Phase 1 study in healthy volunteers showed greater than 60% FXI knockdown following a single 15 mg subcutaneous dose, close to 80% FXI knockdown after a single 50 mg subcutaneous dose, and 85% FXI knockdown three weeks after a single 150 mg subcutaneous dose exceeding our preclinical predictions. CITY-FXI was generally well tolerated by participants dosed in these cohorts. We expect to report additional clinical data from the SAD portion of our Phase 1 study in late 2026 and expect to report data from Part A and Part B SAD cohorts in healthy volunteers and adults with inherited prothrombotic risk factors in mid-2027. Data from the Part C, the multiple-ascending dose (“MAD”) portion of the study in adult participants with history of prior provoked VTE with enduring risk factors, is expected in late 2027. We plan to first evaluate efficacy and safety in thrombotic disease through a Phase 2 trial in TKA, a clinical setting in which patients are at increased risk of postoperative thrombosis. We plan to initiate a Phase 2 trial in TKA in mid-2027 and report data from this trial in early 2028.
Overview of Thromboembolic Diseases
Thromboembolic diseases occur when a blood clot, or thrombus, forms inside veins and arteries. If the thrombus breaks free and travels through the bloodstream to block another vessel, or embolism, the resulting obstruction restricts normal blood flow, preventing vital organs from receiving sufficient oxygen, potentially causing tissue damage or life-threatening emergencies. It is important to distinguish between two related but distinct processes:
•
Hemostasis is the normal physiological response to vascular injury, in which a coordinated sequence of events seals the breach in the vessel wall and arrests bleeding.
•
Thrombosis is a pathological process in which clots form inappropriately within an intact blood vessel, obstructing blood flow and potentially causing injury to downstream tissues.
Thromboembolic diseases represent a major global health burden associated with significant morbidity and mortality through conditions including AFib, myocardial infarction, coronary artery disease, VTE, stroke, and peripheral artery disease, among others. Globally, an estimated 40 to 50 million patients worldwide suffer from these conditions, highlighting a large and underserved patient population.
Secondary strokes lead to devastating outcomes for patients and place a significant burden on the healthcare system, given the high rates of disability, long-term cognitive decline, and mortality. Approximately 795,000 ischemic strokes occur annually in the United States, creating a large population of patients in need of secondary stroke prevention strategies. Currently, approximately 185,000 of these strokes are recurrent and stroke-related costs exceed $56 billion between 2019 and 2020 alone, underscoring the significant clinical and economic value of preventing recurrent events.
AFib, the most common sustained cardiac arrhythmia, is projected to affect approximately 12.1 million people in the United States by 2030. AFib is a leading cause of cardioembolic stroke and is estimated to account for approximately one in seven strokes. Current guidelines recommend oral anticoagulation for AFib patients with a moderate to high stroke risk, a population estimated to encompass approximately 80% of the diagnosed AFib population, or approximately 9.7 million patients in the United States by 2030. Despite the guideline recommendations, many patients remain untreated or under-treated due to contraindications, bleeding concerns, treatment burden, or challenges with long-term adherence. Data from a study of approximately 437,000 patients across nearly 90 U.S. health systems indicate that approximately one in three patients with AFib and an elevated stroke risk score are not receiving any oral anticoagulation. We estimate that the population of DOAC-ineligible and underserved AFib patients in the United States is approximately 3.2 million.
Extended secondary prophylaxis following VTE represents another area of substantial unmet clinical need. VTE affects up to 900,000 people in the United States each year, contributing to an estimated 60,000 to 100,000 deaths annually. Patients who survive an initial VTE event face a meaningful risk of recurrence, with data indicating cumulative recurrence rates of approximately 20% to 30% at five years following a first unprovoked event. Current clinical guidelines recommend considering prolonged anticoagulation in patients for recurrent VTE following an initial event. Despite these recommendations, long-term adherence to currently available anticoagulants is frequently limited by concerns regarding bleeding risk and tolerability, leading many patients to discontinue therapy despite remaining at elevated risk for recurrent thromboembolic events. We estimate that approximately 250,000 to 350,000 patients in the United States remain untreated or under-treated despite having persistent risk factors.
Similarly, cancer-associated thrombosis represents a significant unmet medical need for patients who require antithrombotic protection but are particularly vulnerable to bleeding complications. Patients with active malignancies face an approximately four- to six-fold elevated risk of VTE compared to the general population, and VTE is recognized as the second leading cause of death in cancer patients. Approximately 20% of all VTE events occur in patients with cancer, yet currently available anticoagulants heighten the bleeding risk in oncology patients, particularly those with gastrointestinal or genitourinary cancers. We estimate that the population of cancer patients who require antithrombotic protection in the United States is approximately 200,000.
Current Standard of Care and Unmet Medical Need
There are multiple FDA-approved anticoagulants available for the prevention and treatment of certain types of thromboembolism; however, each class of anticoagulants shares a fundamental mechanistic limitation that constrains their clinical utility. DOACs, vitamin K antagonists, and low-molecular-weight heparins all act on factors within the coagulation cascade that are required for both pathological thrombosis and normal hemostasis. As a result, the anticoagulant activity of these agents is inherently coupled with an increase in bleeding risk.
Elevated bleeding risk precludes some patients from treatment with DOACs, the predominant anticoagulant class, including frail elderly patients, individuals with chronic kidney disease, patients with
active cancer, and those requiring concomitant antiplatelet therapy. Clinical guidelines recognize the challenges of balancing thrombotic risk and bleeding risk in these patient populations. Patient advocacy groups suggest that even seemingly minor adverse reactions, such as bruising or hematuria frequently lead to treatment discontinuation of DOACs. This dynamic may be further exacerbated by the long-term preventative nature of treatment, as the benefit of avoiding a clot is often not directly appreciated by patients in daily life. Additional limitations include dependence on renal clearance, incomplete reversibility, and the need for frequent dosing or laboratory monitoring, each of which can impair adherence. These constraints contribute to undertreatment and dose interruptions. In addition, current anticoagulants are often inadequate for managing thrombosis on artificial surfaces, such as heart valves and catheters, as they lack the mechanistic specificity to prevent device-induced clots without significantly increasing the risk of life-threatening systemic bleeding.
Consequently, there is an urgent and growing need for anticoagulation strategies that can address the bleeding risk associated with current standard of care. We believe that selective inhibition of FXI offers a genetically and clinically supported approach to address pathological thrombosis while preserving normal hemostasis, potentially uncoupling antithrombotic efficacy from bleeding risk and enabling a safer, more effective, and durable treatment option for patients, including those who may be ineligible for current anticoagulants.
Mechanism of Action
The Coagulation Cascade and FXI Biology
FXI is a plasma serine protease that we believe is uniquely positioned in the coagulation cascade where it is key to pathological thrombosis while playing a limited role in normal hemostasis. This distinctive profile makes it an attractive therapeutic target for anticoagulation strategies that aim to prevent clot formation without significantly increasing bleeding risk.
The distinct biology of FXI within the coagulation cascade enables what we believe is a clinically meaningful uncoupling of the body’s protective hemostatic response from the pathological thrombotic process. This separation has not been achieved with current anticoagulants. In healthy coagulation, FXI plays a supporting amplification role: it contributes to clot reinforcement and stability through slow, sustained thrombin generation but is largely dispensable for the rapid, robust hemostatic response required to seal vascular injury (as shown in Figure 1). In normal hemostasis, tissue factor is exposed after vessel injury, triggering strong thrombin generation through the extrinsic pathway, and FXI is not essential for initiating or sustaining clot formation. In contrast, thrombosis occurs primarily within intact vessels and often at sites of plaque rupture, where low levels of tissue factor generate insufficient thrombin to form a clot on their own. Under these conditions, FXI is required to amplify the thrombotic response through a positive feedback loop that drives pathological thrombus formation. Our rationale for selecting FXI as a therapeutic target is because of its central role in pathological thrombosis and limited role in hemostasis.
Figure 1: Differential role of FXI in hemostasis and thrombosis.
*
FIX: Factor IX; FV: Factor V; FVII: Factor VII; FVIII: Factor VIII; FX, Factor X; and TF: Tissue factor.
*
Source: Chan and Weitz, Arterioscler Thromb Vasc Biol. 2023.
This biological uncoupling of thrombosis from hemostasis is supported by human genetic and epidemiological evidence. Individuals with congenital FXI deficiency, including those with severe deficiency (less than 15% of normal activity), rarely experience spontaneous bleeding. When bleeding does occur, it is typically mild and provoked by trauma or surgical procedures, particularly in the oral cavity, nasal passages, and urinary tract. Epidemiological studies further link low FXI activity or antigen levels to reduced risk of thrombotic events. Patients with FXI activity below 50% of normal show significantly lower incidence of ischemic stroke, transient ischemic attack, myocardial infarction, and VTE compared to those with higher FXI levels. Conversely, elevated FXI levels have been associated with increased risk of thrombotic disorders including VTE and ischemic stroke. This natural occurrence in human physiology provides compelling evidence that suppression of FXI can reduce thrombotic risk without compromising baseline hemostatic function. Therapeutic intervention in the FXI pathway for management of thrombosis risk is supported by consistent clinical data, with several therapeutic strategies for FXI inhibition currently under clinical investigation.
DOACs target either Factor Xa or thrombin, which are central nodes of both the hemostatic and thrombosis pathways. By contrast, CITY-FXI targets an upstream amplification step in the coagulation cascade that plays a limited role in physiological hemostasis and is therefore intended to provide antithrombotic efficacy with a substantially safer bleeding profile. FXI-targeted inhibition may be suitable for prolonged use and in patients for whom current anticoagulants are contraindicated or associated with prohibitive bleeding risk, such as the elderly, those with chronic kidney disease, or oncology patients undergoing chemotherapy.
Novel Approaches
FXI inhibition has emerged as one of the most actively pursued areas of cardiovascular drug development, reflecting the scientific consensus that upstream targeting of the thrombosis coagulation pathway may enable the separation of antithrombotic efficacy from bleeding liability. There are currently no approved therapeutics that directly target FXI, however, several candidates are in clinical development, including milvexian, asundexian, abelacimab, REGN7508, REGN9933, and CTX-611. Phase 2 and Phase 3 data for other FXI agents have established proof-of-concept by reducing thrombotic risk without elevating bleeding risk.
The majority of clinical-stage FXI inhibitors fall under the categories of small molecules and antibodies, though both are mechanistically distinct from FXI knockdown by RNAi and carry limitations compared to CITY-FXI’s mechanism of action (as shown in Figure 2). CITY-FXI reduces hepatic production of FXI, resulting in depletion of the circulating FXI protein and broad suppression of FXI-mediated activity. By reducing the protein itself rather than inhibiting a specific functional site, this approach is designed to inhibit all known FXI functions and more closely recapitulates the protective phenotype observed in individuals with genetically reduced FXI levels.
In contrast, small-molecule FXI inhibitors generally target the catalytic activity of activated FXI (FXIa) and therefore inhibit only a subset of FXI-mediated biological functions. As a result, these agents may not fully block non-catalytic FXI interactions that contribute to thrombin generation and coagulation pathway amplification.
Similarly, anti-FXI antibodies are designed to inhibit specific FXI functions, including zymogen activation (the conversion of an inactive precursor into its active form) or enzymatic activity, depending on their binding site and mechanism of action. The degree of pathway inhibition varies based on target engagement and binding affinity, and is often relatively short lived.
We believe that the broader and more durable suppression of FXI achieved with CITY-FXI offers the potential for more complete pathway inhibition and favorable efficacy, safety, and dosing.
Figure 2: FXI knockdown is categorically different from small molecule and antibody inhibition.
SOC = Standard of Care
Overview of CITY-FXI
CITY-FXI is designed to specifically and effectively suppress FXI at the mRNA level, resulting in near-complete elimination of the protein. CITY-FXI is a City-engineered GalNAc-conjugated siRNA therapeutic candidate designed to selectively engage the asialoglycoprotein receptor (“ASGPR”) on hepatocytes for intracellular delivery. Once internalized, CITY-FXI is designed to suppress hepatic expression of F11 mRNA (the messenger RNA transcript encoding the FXI protein) via RNAi, resulting in highly selective and durable suppression of hepatic FXI synthesis.
The clinical viability of GalNAc-siRNA therapy has been demonstrated for several approved RNAi drugs, including LEQVIO, AMVUTTRA, GIVLAARI, OXLUMO, RIVFLOZA, QFITLIA, and TRYNGOLZA. CITY-FXI combines a genetically and clinically supported target with a potent and specific City-engineered RNAi trigger and a proven GalNAc delivery approach, with the aim of providing a potent, specific, and durable antithrombotic effect while preserving normal hemostatic function.
Preclinical Data
CITY-FXI has demonstrated compelling preclinical data supporting its development as an anticoagulation treatment targeting FXI. In NHPs, a single subcutaneous dose of CITY-FXI produced sustained, dose-dependent plasma FXI suppression exceeding 95% at 10 mg/kg, with maximal suppression typically occurring between Days 21 and 28. This translated into durable pharmacodynamic activity, as reflected by prolongation of activated partial thromboplastin time (“aPTT”), a measure of blood clotting time, consistent with the expected effect of FXI suppression on the thrombosis pathway. Maximum mean aPTT fold increases over baseline reached 1.28, 1.46, and 1.67 at 1 mg/kg, 3 mg/kg, and 10 mg/kg subcutaneous, respectively, with effects lasting up to 13 weeks (as shown in Figure 3).
Figure 3: 95% FXI knockdown in NHP with prolonged clotting time upon single subcutaneous dose of CITY-FXI.
In the preclinical studies of CITY-FXI, potent, selective, and durable RNAi-mediated suppression of hepatic F11 mRNA expression and consequently circulating FXI protein was observed, as well as favorable pharmacokinetic and pharmacodynamic profiles, a lack of off-target transcriptional or genotoxic effects, and a generally well-tolerated preclinical profile that together established adequate safety margins and supported progression to first-in-human (“FIH”) dosing.
Clinical Data
The first clinical trial of CITY-FXI is a Phase 1, dose-ranging, proof-of-biology trial designed to evaluate the safety, pharmacokinetics, and pharmacodynamics in three populations:
•
Part A: SAD trial in adult healthy volunteers, who will receive a single dose of CITY-FXI in one of three dose cohorts: 15 mg, 50 mg, or 150 mg. Each cohort will include eight participants; six will receive CITY-FXI and two will receive placebo.
•
Part B: SAD trial in adult participants with inherited prothrombotic risk factors, such as Factor V Leiden (“FVL”) or the prothrombin G20210A mutation, who will receive CITY-FXI in one of up to three cohorts at higher dose levels starting at 300 mg, and potentially increasing to 600 mg and 900 mg, subject to approval by a safety review committee. Each cohort will include eight participants; six will receive CITY-FXI and two will receive placebo.
•
Part C: Adult participants with a history of prior provoked VTE with enduring risk factors, with up to three MAD cohorts to be informed by results of Part B. Each cohort will include nine participants; six will receive CITY-FXI and three will receive placebo.
At the time of this report, participants in all three cohorts of Part A have been dosed. In each cohort, six healthy adults received a single subcutaneous dose of CITY-FXI (15 mg for cohort 1, 50 mg for cohort 2, and 150 mg for cohort 3). FXI knockdown at an average of 61% was observed in cohort 1, an average of 79% FXI knockdown was observed in cohort 2, and at three weeks after dosing, an average of 85% FXI knockdown was observed in cohort 3, demonstrating CITY-FXI’s robust, dose-dependent response (as shown in Figure 4). CITY-FXI was generally well tolerated in all participants. No serious adverse events were observed and no stopping rules were met.
Figure 4: FXI knockdown in healthy volunteers upon single dose of CITY-FXI.
Data: Mean ± SD (n=6); as of August 14, 2026 data cutoff.
Further, durable FXI suppression is projected with repeated CITY-FXI dosing. We have generated a pharmacokinetic/pharmacodynamic (“PKPD”) model based on the FXI suppression observed in NHPs following a single subcutaneous dose of CITY-FXI as well as the clinical experience of other GalNAc-conjugated siRNAs. Based on results from the Part A, cohort 1 and cohort 2 of the Phase 1 trial, our PKPD model projects sustained FXI suppression ≥98% with once every three months (“Q3M”) dosing and ≥94% with once every six months (“Q6M”) dosing, each at a dose of 300 mg (as shown in Figure 5).
Figure 5: Durable FXI suppression projected with repeat CITY-FXI dosing. Sustained ≥98% following Q3M and ≥94% following Q6M 300 mg of CITY-FXI based on initial clinical data.
Overall, in both the clinical and preclinical studies, we observed potent, selective, and durable RNAi-mediated suppression of FXI, favorable pharmacokinetic and pharmacodynamic profiles, lack of off-target transcriptional effects, and a generally well-tolerated profile. We expect to report additional data from the SAD portion of our Phase 1 trial in late 2026 and expect to report data from Part A and Part B SAD cohorts in healthy volunteers and adults with inherited prothrombotic risk factors in mid-2027. Data from the Part C, the MAD portion of the study in adult participants with history of prior provoked VTE with enduring risk factors, is expected in late 2027.
We plan to first evaluate efficacy and safety in thrombotic disease through a Phase 2 trial in TKA, a clinical setting in which patients are at increased risk of postoperative thrombosis. The data from this TKA study will be used to support initiation of Phase 3 studies. We plan to initiate the Phase 2 trial in TKA in mid-2027 and to report data from this study in early 2028.
Given the significant market opportunity for CITY-FXI, the large-scale Phase 3 clinical trials expected to be required for regulatory approval, and the substantial commercialization resources required to support a potential launch, we expect to seek a collaboration partner for the asset prior to initiating Phase 3 development.
CITY-RBP4
CITY-RBP4 is a City-engineered GalNAc-conjugated siRNA therapeutic candidate designed to target RBP4 mRNA for the treatment of retinal diseases characterized by the toxic accumulation of vitamin A byproducts. We are initially developing CITY-RBP4 to treat Stargardt disease, an inherited retinal disorder caused by mutations in the ABCA4 gene, which encodes the retinal-specific ATP-binding cassette transporter A4 responsible for clearing vitamin A byproducts from the retina. When ABCA4 function is impaired, these byproducts accumulate as toxic bisretinoids, leading to progressive retinal degeneration.
Inhibition of RBP4 is a clinically supported mechanism for Stargardt disease, and to our knowledge CITY-RBP4 is the first and only RNAi therapy in development aimed at inhibiting RBP4. By targeting and suppressing RBP4 mRNA in the liver, CITY-RBP4 aims to reduce vitamin A delivery to the eye and slow the progression of retinal damage associated with the disease. In a preclinical mouse model of Stargardt disease, the suppression of RBP4 through RNAi normalized ocular A2E, a major toxic vitamin A byproduct, to wild-type levels at therapeutically relevant doses. In a preclinical study in NHPs, following treatment with CITY-RBP4, we observed approximately 90% suppression of circulating vitamin A and RBP4 at dose levels of 3 mg/kg and 10 mg/kg.
We filed an application for CITY-RBP4 in mid-2026 to initiate a Phase 1 clinical study in late 2026 to evaluate its safety, tolerability, pharmacokinetics, and pharmacodynamics in healthy volunteers and Stargardt patients. We expect to report initial data from the Phase 1 clinical trial in mid-2027 and data from the SAD and MAD cohorts in healthy volunteers and Stargardt patients in early 2028. We are also exploring CITY-RBP4 in adjacent indications, including GA secondary to age-related macular degeneration. We expect to initiate a Phase 2 clinical trial for CITY-RBP4 in GA in late 2027.
Overview of Stargardt Disease Type 1
Stargardt disease is an inherited macular degenerative disease and the leading inherited cause of central vision loss in children and young adults. More than 30,000 people in the United States are diagnosed with Stargardt disease, and some studies suggest that the prevalence is even higher, indicating there could be a significant number of undiagnosed patients who emerge upon FDA approval of a treatment.
Stargardt disease is an autosomal recessive disorder caused by mutations in the ABCA4 gene, causing progressive and irreversible vision loss usually beginning in early childhood, with the majority of patients becoming legally blind in their late twenties. Onset typically occurs during critical periods of educational, vocational, and psychosocial development and results in increasing limitations in mobility, reading, and facial recognition. The burden of disease extends well beyond visual impairment: Patient-reported rates of depression and anxiety significantly exceed those of the general population and vision loss frequently impairs personal autonomy, workforce participation, and long-term employment.
Current Standard of Care and Unmet Medical Need
There are no currently approved treatments for Stargardt disease. Consequently, patient management is limited to lifestyle counseling and vision rehabilitation. Diagnosis ranges from early childhood to early adulthood depending on the mutation in ABCA4 and is often delayed by several years relative to symptom onset. Patients are generally advised to avoid vitamin A supplementation, smoking, and excessive exposure to high-intensity light, but these measures do not prevent the progressive vision loss associated with the disease. There is an urgent need for disease-modifying therapeutics that address the underlying pathophysiology of Stargardt disease and slow the progression of irreversible retinal degeneration.
The Visual Cycle and Role of RBP4 in Stargardt Disease
In Stargardt disease, mutations in the ABCA4 gene impair the retina’s ability to process and clear vitamin A-derived byproducts, resulting in the accumulation of toxic, vitamin A bisretinoids, including A2E, within retinal pigment epithelium cells. These compounds drive retinal degeneration through multiple mechanisms, including cellular toxicity, oxidative stress, and complement activation. Over time, this progressive accumulation of these toxic byproducts leads to retinal dysfunction, cell loss, and irreversible vision impairment.
CITY-RBP4 is designed to address this underlying pathology by reducing hepatic production of RBP4, the primary transporter responsible for delivering vitamin A from the liver to the eye. By reducing delivery of vitamin A to the retina, CITY-RBP4 aims to limit the formation and accumulation of toxic bisretinoids, preserve retinal cells, and slow the progression of vision loss associated with Stargardt disease.
Novel Approaches for Stargardt Disease
There are currently no approved therapies for Stargardt disease. Current RBP4 therapies in development include tinlarebant, which has demonstrated in clinical trials that reducing circulating RBP4 levels is a potentially effective strategy for treating Stargardt disease. Other non-RBP4 approaches in clinical development include vitamin A analogs and gene therapies.
The vitamin A analog gildeuretinol blocks the formation of toxic bisretinoids by diluting the body’s natural vitamin A with deuterated vitamin A, and by doing so, slowing toxic bisretinoid aggregation. This approach requires clinical doses at levels well above the recommended daily dose of vitamin A, posing a risk for hypervitaminosis A.
Most gene therapies aim to restore ABCA4 expression to suppress toxic metabolite accumulation; however, these approaches remain largely in early development, with unresolved questions regarding efficacy and long-term safety. In addition, they require highly invasive local injections that carry meaningful procedural risk.
Collectively, these limitations highlight the need for therapeutic approaches with the potential for durable efficacy, better safety, and more convenient dosing.
Overview of CITY-RBP4
CITY-RBP4 is a City-engineered GalNAc-conjugated siRNA therapeutic candidate specifically designed to suppress RBP4 mRNA in the liver, with the goal of reducing retinol delivery to the eye and decreasing the formation of toxic bisretinoids that lead to vision loss. We believe CITY-RBP4 combines a clinically supported mechanism with a potent and specific RNAi therapeutic. By degrading RBP4 mRNA before the protein is produced, CITY-RBP4 has the potential to achieve deeper and more durable knockdown than small molecules, which inhibit RBP4 only after the protein has been produced. This durable suppression avoids the peak-and-trough oscillation in drug exposure characteristic of small molecules, instead providing steadier, more favorable pharmacology that we believe will translate into greater patient benefit. Because CITY-RBP4 is designed to be delivered directly to the liver rather than being systemically distributed, it has the potential to reduce off-target effects in non-target tissues. In addition, unlike small molecules, which require consistent daily dosing, CITY-RBP4 is being studied as a potentially quarterly to biannual subcutaneous injection, which may improve adherence, particularly in adolescent patients during a critical time of disease progression.
Preclinical Data
In the ABCA4-/- mouse model, a well-established preclinical model of Stargardt disease, treatment with an siRNA targeting mouse RBP4 dosed at 3 mg/kg showed reductions in plasma RBP4 and retinol levels, and levels of ocular A2E, the primary component of toxic vitamin A bisretinoids, were completely normalized at a therapeutically relevant dose. Furthermore, the reduction in A2E was dose-dependent, suggesting that the target engagement of RBP4 is specific (as shown in Figure 6).
Figure 6: siRNA-mediated RBP4 suppression observed to reduce circulating vitamin A and normalize ocular A2E in the ABCA4−/− Stargardt disease mouse model.
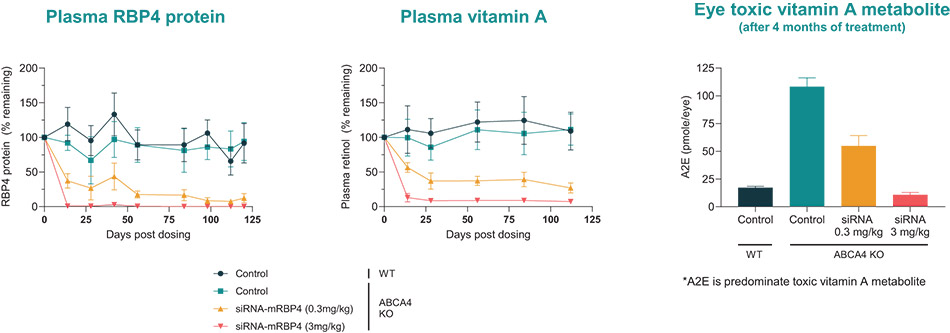
Sustained suppression of plasma RBP4 with correlating suppression of plasma vitamin A was also observed in NHPs. Maximum RBP4 and retinol suppression was observed between Day 15 and Day 29 following a single subcutaneous administration of CITY-RBP4, with suppression occurring in a dose-dependent manner across all dose levels. At the moderate dose of 3 mg/kg, we observed approximately 90% RBP4 and vitamin A suppression (as shown in Figure 7). Further, at the highest subcutaneous dose of 10 mg/kg, RBP4 and retinol mean suppression both exceeded 90%, peaking on Day 29, with substantial suppression up to Day 90. A near 1:1 correlation was observed between RBP4 and vitamin A levels in NHPs, supporting that RBP4 is the main carrier for vitamin A delivery to peripheral tissues, including the eye (as shown in Figure 7). This biological relationship provides strong mechanistic support for RBP4 as a compelling therapeutic target for reducing vitamin A delivery in Stargardt disease.
Figure 7: >90% RBP4 and vitamin A suppression observed with one subcutaneous dose of CITY-RBP4. Plasma RBP4 and vitamin A levels exhibit near 1:1 correlation in NHP.
In the preclinical studies of CITY-RBP4, potent, selective, and durable RNAi-mediated suppression of hepatic RBP4 mRNA was observed, with consequent suppression of circulating RBP4 protein and retinol. The data also showed favorable pharmacokinetic and pharmacodynamic profiles, and a lack of off-target transcriptional or genotoxic effects.
Clinical Development Plan
We filed an application for CITY-RBP4 in mid-2026 to initiate a Phase 1 clinical study in late 2026. We expect the Phase 1 study to be a multicenter, randomized, double-masked, placebo-controlled study evaluating the safety, tolerability, pharmacokinetics, and pharmacodynamics of CITY-RBP4 administered subcutaneously in healthy volunteers and patients diagnosed with Stargardt disease. We expect the study to consist of SAD and MAD cohorts. We expect to report initial data from the Phase 1 trial in mid-2027 and data from the SAD and MAD cohorts in healthy volunteers and Stargardt patients in early 2028.
Indications Beyond Stargardt disease — Geographic Atrophy (GA) Secondary to Age-Related Macular Degeneration
GA is an advanced form of age-related macular degeneration that shares pathology with Stargardt disease given the accumulation of toxic vitamin A bisretinoids in the eye. In GA, the progressive, irreversible loss of retinal cells leads to expanding patches of vision loss in the central retina. GA affects approximately one million people in the United States and approximately five million people globally, yet there are only two approved therapies, both of which require frequent and invasive intravitreal injections. Despite recent approvals, a significant unmet medical need persists, as available therapies achieve only a modest slowing of lesion growth. GA and Stargardt disease share a common pathway of vitamin A-induced retinal toxicity; therefore, we believe CITY-RBP4 has strong therapeutic potential to mitigate progressive retinal degeneration in both indications. To support this expansion, we are planning to evaluate CITY-RBP4 in healthy volunteers aged 60 or older, aligning with the typical age of GA onset. We expect to initiate a Phase 2 clinical trial for CITY-RBP4 in GA in late 2027.
CITY-TFR2
CITY-TFR2 is a City-engineered GalNAc-conjugated siRNA therapeutic candidate designed to reduce TFR2 expression, lower hepcidin levels, and restore iron availability for red blood cell production. We are initially developing CITY-TFR2 for myelofibrosis-associated anemia, where anemia is a major driver of morbidity and transfusion dependence, while also exploring broader applications in anemia associated with chronic kidney disease and chronic inflammatory disorders, including inflammatory bowel disease, where elevated hepcidin and iron restriction also contribute to disease burden. An IIT for CITY-TFR2 is expected to begin in China for the treatment of anemia in myelofibrosis patients in late 2026, with a goal of demonstrating clinical proof-of-concept. Under an IIT study agreement entered into in July 2026, we have contracted with Overland, a biotechnology company with operations in the United States and China, to provide clinical and operational support for the IIT. Pursuant to the agreement, we will provide support for the IIT, supply CITY-TFR2, and fund covered study-related services and expenses through milestone payments, and Overland, as local study sponsor, will provide clinical and operational support in connection with the IIT, including supporting site and investigator selection, administrative submissions, project management, insurance and clinical monitoring, and, subject to our prior approval, will enter into separate agreements with the applicable investigators, hospitals and research institutions in China. The IIT will be conducted by investigators at participating medical institutions, with clinical and operational support and coordination provided by Overland. While the IIT is not one of our traditional company-sponsored clinical trials, subject to applicable law and the applicable agreements with the investigators and medical institutions, data and results generated from the IIT will be provided to us. Although data from the IIT may support our development program for CITY-TFR2, including by providing initial clinical proof-of-concept data and informing our planned Phase 1/2 clinical trial, there can be no assurance that the FDA or other regulatory authorities will accept such data to support future regulatory submissions or marketing approval. We expect to report data from the IIT in mid-2027.
We also plan to file an IND or IND-equivalent application in late 2026, initiate a Phase 1/2 clinical trial for CITY-TFR2 in early 2027 and report data from the Phase 1/2 study in late 2027. Assuming positive results from the IIT and Phase 1/2 studies, we plan to initiate a Phase 2 study in an additional indication of anemia of chronic disease in early 2028.
Overview of Anemia of Chronic Disease
Anemia of chronic disease comprises up to 40% of all anemias and affects more than 5 million people in the United States. Anemia of chronic disease is a common complication of chronic inflammatory and malignant disorders, including myelofibrosis, chronic kidney disease, and inflammatory bowel disease, and is characterized by a shortage of healthy red blood cells, or erythrocytes, available to carry oxygen to the body’s tissues, causing persistent fatigue, reduced exercise tolerance, and diminished quality of life. In many patients, anemia progresses to blood transfusion dependence, which is highly burdensome to patients and the healthcare system, and can lead to serious complications such as iron overload. Anemia can be broadly classified into two types: iron-depleted and iron-restricted anemia.
Unlike anemia caused by iron deficiency, where iron stores are depleted, iron-restricted anemia occurs when iron is present but unavailable for erythropoiesis, or red blood cell production. Anemia of chronic disease is a type of iron-restricted anemia. A hallmark of anemia of chronic disease is inflammation-driven iron restriction, where inflammatory cytokines, particularly interleukin-6 (“IL-6”), stimulate production of the hormone hepcidin, the central regulator of systemic iron homeostasis. Elevated hepcidin restricts the availability of iron needed for erythropoiesis by preventing iron release from cellular storage sites and limiting iron absorption. As a result, many patients develop iron-restricted anemia despite having adequate total body iron stores.
Although the mechanisms contributing to anemia differ among these conditions, elevated hepcidin and the resulting iron restriction contribute to anemia across these diseases. Therapies capable of restoring iron availability may have broad utility across multiple diseases associated with iron-restricted anemia.
Current Standard of Care and Unmet Medical Needs
Iron-restricted anemia remains a major source of morbidity across myelofibrosis, chronic kidney disease, and chronic inflammatory disorders, including inflammatory bowel disease, yet current treatment options are limited by modest efficacy, significant safety concerns, and heavy treatment burden.
We are initially developing CITY-TFR2 for anemia of myelofibrosis. Myelofibrosis is a rare, chronic cancer of the bone marrow in which the marrow becomes progressively scarred and loses its capacity to produce blood cells normally. There are an estimated 25,000 patients in the United States with myelofibrosis. Studies estimate that more than 85% of patients develop anemia. Treatment options include ESAs, immunomodulatory agents including thalidomide, lenalidomide, and pomalidomide (often administered with prednisone), and androgen therapies such as danazol. Responses to these therapies are often limited or transient, and many patients ultimately become dependent on blood transfusions. In addition, treatments indicated for myelofibrosis that modulate the JAK-STAT pathway are frequently associated with worsening anemia, resulting in treatment discontinuations or patients being treatment ineligible. Consequently, there is a significant unmet medical need for effective treatments of anemia in myelofibrosis.
CITY-TFR2 also has the potential to treat anemia in additional inflammatory indications, such as chronic kidney disease and chronic inflammatory disorders, including inflammatory bowel disease, by targeting the shared pathway of hepcidin-mediated iron restriction. Chronic kidney disease affects an estimated 37 million adults in the United States, approximately 4.8 million of whom develop anemia as kidney function declines. Current treatments include ESAs, HIF-PH inhibitors, intravenous iron and blood transfusions. ESAs and HIF-PH inhibitors are associated with significant safety concerns, including boxed warnings, and many patients continue to experience persistent anemia despite treatment.
Inflammatory bowel disease, comprising Crohn’s disease and ulcerative colitis, affects an estimated three million adults in the United States, with 21-27% of patients developing anemia over the course of their disease. While iron supplementation can address iron deficiency, inflammation-driven iron restriction may persist despite iron repletion, leaving many patients with ongoing anemia and limited treatment options.
By targeting hepcidin-mediated iron restriction, a shared driver of anemia across these diseases, CITY-TFR2 is designed to address a common mechanism underlying anemia of chronic disease and has the potential to be broadly applicable across multiple indications.
TFR2 Biology and the Role of Hepcidin in Iron Homeostasis
Under normal conditions, the liver-derived hormone hepcidin acts as a molecular thermostat, rising when iron levels are high to limit iron absorption, and falling when iron is low to mobilize stores for red blood cell production. In anemia of chronic disease, the overproduction of inflammatory cytokines including IL-6, triggers the overproduction of hepcidin, trapping iron inside cells (as shown in Figure 8). This creates a state of iron restriction, severely limiting red blood cell production despite abundant systemic iron stores.
Figure 8: Anemia of chronic disease short-circuits the body’s iron thermostat: iron homeostasis requires feedback between liver and bloodstream through hepcidin.
TFR2 is a key iron-sensing protein expressed on liver cells that helps regulate hepcidin and systemic iron availability. TFR2 binds to holo-transferrin (iron-bound transferrin), thereby linking hepcidin production to transferrin saturation and systemic iron availability.
Human genetic evidence supports TFR2’s role as a modulator of systemic iron balance, as complete loss-of-function mutations in TFR2 are associated with hereditary hemochromatosis type 3, a disorder characterized by reduced hepcidin levels and increased intestinal iron absorption, resulting in systemic iron overload. However, clinically meaningful manifestations typically emerge only after decades of elevated iron levels. Individuals with partial loss-of-function of TFR2 generally do not develop iron overload. These observations support the approach of pharmacologically modulating TFR2 to achieve controlled improvements in iron regulation.
CITY-TFR2 & Other Novel Approaches for Anemia of Chronic Disease
CITY-TFR2 is a TFR2-targeting, City-engineered GalNAc-conjugated siRNA therapeutic candidate designed to reduce the expression of TFR2, thereby lowering hepcidin levels and increasing iron availability for red blood cell production. Recent third-party clinical data with an anti-hemojuvelin (“HJV”) antibody, selcodebart (DISC-0974), demonstrated meaningful increases in hemoglobin in patients with myelofibrosis-associated anemia, supporting the therapeutic potential of targeting the hepcidin pathway.
We believe TFR2 represents a particularly attractive target because it directly senses circulating iron through its interaction with holo-transferrin and therefore serves as a physiologically relevant regulator of hepcidin production. In contrast, HJV is more closely associated with sensing tissue and intracellular iron stores. We believe TFR2 also offers advantages in inflammatory disease settings. While HJV expression is downregulated by inflammation, potentially limiting the effectiveness of approaches that depend on HJV biology, TFR2 remains a relevant regulator of hepcidin in inflammatory states, which we believe makes it a compelling therapeutic target in diseases characterized by chronic inflammation. Based on the results of our preclinical studies, we believe CITY-TFR2 also has the potential to provide durable target suppression and infrequent dosing, which may translate into greater adherence and sustained benefit relative to antibody-based approaches.
Preclinical Data
Preclinical findings support TFR2 as a promising therapeutic target for addressing the iron restriction and impaired red blood cell production that characterize anemia of chronic disease. In a diet-induced mouse model of anemia, suppression of TFR2 with an siRNA targeting mouse TFR2 resulted in an increase in transferrin saturation (“TSAT”), and mobilization of iron stores to the periphery, that resulted in maintenance of the hematocrit at normal baseline levels (as shown in Figure 9). In contrast, mice treated with saline showed a reduction in TSAT and a hematocrit lower than baseline.
Figure 9: Suppression of TFR2 in a mouse model of anemia maintained normal hematocrit.
Following subcutaneous dosing of CITY-TFR2 in NHPs, we observed robust and sustained knockdown of TFR2 mRNA and concomitant increases in circulating iron, TSAT, and mean corpuscular hemoglobin (“MCH”), a measure of hemoglobin content per red blood cell. In a two-dose study in NHPs, subcutaneous dosing of CITY-TFR2 led to TFR2 mRNA reduction of over 80% after the first dose at dose levels of 1 mg/kg and 3 mg/kg and 89% after the second dose at 3 mg/kg (as shown in Figure 10). CITY-TFR2 also increased serum iron, TSAT, and MCH in NHPs after both doses with durability lasting 98 days after the second dose (as shown in Figure 11).
Figure 10: Robust and durable knockdown of TFR2 mRNA observed in NHPs.
*
TFR2 mRNA measured from sparse sampling of liver biopsies taken across NHPs (N=1 per timepoint through day 121). All CITY-TFR2 animals had a liver biopsy on the last day (day 176).
Figure 11: Suppression of TFR2 increased serum iron, TSAT, and MCH in NHPs.
*
All data represented as Mean ± SEM
In the preclinical studies of CITY-TFR2, a profile characterized by potent, selective, and sustained RNAi-mediated silencing of TFR2 mRNA was observed, as well as a subsequent increase in serum iron, TSAT, and MCH. These physiological responses were coupled with highly favorable pharmacokinetic and pharmacodynamic properties and negligible off-target transcriptional activity.
Clinical Development Plan
CITY-TFR2 is being developed as a potential treatment for multiple diseases characterized by inflammation-driven iron restricted anemia. We believe the underlying mechanism may be applicable across anemias in myelofibrosis, chronic kidney disease, and chronic inflammatory disorders, including inflammatory bowel disease. Our initial clinical focus is myelofibrosis, where anemia represents a major unmet medical need, and where hepcidin-mediated iron restriction is believed to play a significant role in disease burden and hepcidin downregulation has been clinically demonstrated as a potentially effective treatment approach. Myelofibrosis is associated with pro-inflammatory pathways that contribute
directly and indirectly to suppression of erythropoiesis while also interacting with systemic iron regulation. Therapeutic modulation of JAK-STAT signaling improves splenomegaly and symptoms in many myelofibrosis patients but is frequently associated with worsening anemia. In some patients, JAK inhibitors cannot be prescribed due to anemia. Consequently, there is a significant unmet medical need for therapies that restore effective red blood cell production, provide sustained improvements in hemoglobin, reduce transfusion burden, and more directly address the interplay between inflammation-driven iron restriction and red blood cell production.
We plan to initiate the clinical development of CITY-TFR2 through an IIT in China, with a goal of establishing clinical proof-of-concept. The IIT is expected to include SAD and MAD cohorts in patients with myelofibrosis associated anemia. We anticipate initiating the IIT in late 2026, with clinical data expected in mid-2027. In addition to the IIT, we plan to file an IND or IND-equivalent application for a Phase 1/2 clinical trial in late 2026, with trial initiation anticipated in early 2027. The Phase 1/2 trial, which will include healthy volunteers and patients with myelofibrosis-associated anemia, is expected to further evaluate and extend pharmacodynamic and tolerability data across a broader myelofibrosis patient population. We expect to report data from the Phase 1/2 study in late 2027. Data generated from the myelofibrosis program will help inform future development in additional indications. Assuming positive results from the IIT and Phase 1/2 studies, we plan to initiate a Phase 2 study in an additional indication of anemia of chronic disease in early 2028.
Our Approach: Potential for RNAi Therapeutics and Our Platform
The City Platform
The City Platform is the engine behind every therapeutic we develop. Our City Platform is designed to integrate three core capabilities: RNAi trigger engineering, novel delivery conjugates that allow for tissue selectivity, and target discovery and selection enabled by our insights into human genetics. Critical to every RNAi medicine is the trigger that harnesses the cellular RNAi machinery to silence a disease-relevant gene. We believe our expertise enables us to engineer the trigger for high potency and specificity, and our novel cityRNA triggers are a differentiated approach to unlock extrahepatic delivery. Our proprietary AI/ML-enabled trigger design capability uses a large RNAi activity dataset to predict high-potential trigger molecules in silico, accelerating design cycles from months to weeks. We conjugate our triggers with tissue-targeting ligands for precise delivery to cell types in and beyond the liver, starting with our proprietary LRP1 ligand, which is designed to enable delivery to ocular, CNS, and adipose tissues.
Our pipeline programs are selected through a genetics-led target discovery and selection capability that leverages proprietary insights from human biology and other sources of substantiation, including clinical data. We use this capability to identify differentiated genetic targets where RNAi may be able to confer a durable therapeutic advantage. By uniting these capabilities in a single platform, we aim to systematically create and develop potentially best-in-class therapeutics, supporting our goal of filing one to two new IND or IND-equivalent applications per year. The filing of an IND or IND-equivalent application, or the clearance of any such application by the FDA or a comparable foreign regulatory authority, is not indicative of the future success of any of our product candidates and does not increase the likelihood that any of our product candidates will receive regulatory approval. Given our early stage of development, it will take many years before we complete clinical development of and receive regulatory approval for any of our product candidates, if ever.
Figure 12: The City Platform: trigger engineering at the core, with novel delivery conjugates and target discovery in support.
Trigger Engineering
The RNAi trigger is fundamental to the performance of every RNAi medicine and is a double-stranded RNA that programs the cell’s RNA induced silencing complex (“RISC”) machinery to cleave a target mRNA and induce gene silencing. RNAi trigger engineering is the cornerstone of our City Platform, leveraging our deep expertise in canonical siRNA therapeutics, our next-generation cityRNA triggers, and our proprietary AI/ML molecular-design capability.
Our team of RNAi pioneers bring deep, first-hand expertise in the design, chemical modification, screening, and engineering of RNAi trigger molecules to create RNAi therapeutics. This expertise stems from our contributions to the discovery or development of seven of the eight currently approved RNAi therapeutics. We believe that our combined expertise positions us to directly compete in the large and growing market for liver-directed RNAi therapies with the goal of developing potentially best-in-class siRNAs. Our team’s collective experience is also the foundation on which cityRNA trigger molecules are built — the same design ingenuity and discipline, chemistry and screening infrastructure, applied to a structurally new class of RNAi molecule.
CityRNA — Our Proprietary Next-Generation RNAi Trigger
While there is substantial opportunity for liver-directed RNAi therapeutics, we believe the defining opportunity for the future of RNAi therapeutics lies beyond the liver. The liver is uniquely permissive to RNAi because hepatocytes abundantly express ASGPR, the receptor for the GalNAc ligand, and are readily accessible from the bloodstream through the liver’s highly permeable vasculature. However, the majority of human diseases arise in tissues beyond the liver, which lack a comparably efficient uptake pathway and present far more restrictive extracellular, endothelial, and cellular barriers. In settings where delivery is constrained, the intrinsic molecular properties of the trigger become decisive in successfully developing an RNAi therapeutic. cityRNA is our proprietary trigger, engineered specifically to perform where conventional approaches fall short, enabling expansion to extrahepatic tissues. cityRNA trigger molecules are differentiated based on six advantages:
1.
Improved biophysical properties. cityRNA trigger molecules are approximately 30% shorter than canonical siRNAs and carry roughly one-third fewer negative charges. These structural features are well suited to the biological realities of extrahepatic tissues, where molecular size and charge strongly influence interactions with plasma proteins, biological barriers, and cellular membranes that ultimately determine tissue distribution and cellular uptake. By reducing electrostatic repulsion at the cell surface, the lower charge density may also
enhance penetration through dense extracellular matrices. Most importantly, subcellular imaging suggests that cityRNAs are inherently more efficient at escaping endosomes, allowing more of the delivered payload to reach the cytoplasm and engage RISC (as shown in Figure 13). This increased efficiency could be transformative for extrahepatic RNAi, where delivery is inherently constrained and the therapeutic impact depends on extracting maximal activity from every molecule that enters the cell.
Figure 13. Reduced co-localization of cityRNA-GalNAc with the late endosomal marker Rab7A compared to siRNA-GalNAc.
2.
Dual AGO2 and AGO3 utilization and higher catalytic turnover. cityRNA trigger molecules are designed to engage the RNAi-silencing machinery with increased productivity. Canonical siRNAs function almost exclusively through the Argonaute 2 (“AGO2”)-RISC. When loaded into Argonaute 3 (“AGO3”)-RISC, siRNA becomes trapped in a dead-end complex that sequesters payload without contributing to silencing. Owing to its shorter length, cityRNA can convert AGO3-RISC from a molecular sink into an active, cleavage-competent enzyme, productively engaging both AGO2 and AGO3 (as shown in Figure 14), which are both universally expressed in all tissues. To our knowledge, we are the only company developing RNAi triggers that productively engage both pathways. In addition, cityRNA-loaded RISC improves catalytic turnover by releasing cleaved mRNA more efficiently, freeing it for the next round of mRNA cleavage (as shown in Figure 15). In delivery-constrained settings, a molecule that achieves more productive silencing per copy delivered offers a meaningful practical advantage.
Figure 14. cityRNA enables productive cleavage in both AGO2-RISC and AGO3-RISC, whereas siRNA activity is largely restricted to AGO2-RISC.
Figure 15. cityRNA enhances AGO2-RISC catalytic turnover, producing more rapid and greater cumulative product formation than siRNA.
3.
Improved specificity. Head-to-head preclinical data generated to date demonstrate an equivalent or superior off-target profile for cityRNA trigger molecules versus canonical siRNA triggers bearing identical seed sequences, the region of the trigger responsible for target recognition. We believe that a cleaner off-target profile may widen the therapeutic index, a particularly valuable property in sensitive extrahepatic tissues such as the eye and CNS, where delivery efficiency is constrained and off-target activity could carry greater tolerability risk. cityRNA’s intrinsic potency may lower the effective dose required for target knockdown, further reducing the off-target burden and potentially improving tolerability.
4.
Increased design flexibility. cityRNA trigger molecule’s distinct structural properties may expand the landscape of viable trigger sequences that can be explored for each target. This increases the probability of identifying triggers that are simultaneously potent, cross-species homologous, and differentiated. A larger proprietary trigger-sequence estate may allow us to identify RNAi trigger sequences that are novel to existing siRNA programs.
5.
Lower cost of goods and more scalable manufacturing. cityRNA trigger molecule’s approximately 30% shorter oligonucleotide length simplifies synthesis and reduces raw-material requirements relative to the canonical 21-25 nucleotide duplex. We believe this advantage will scale as our pipeline grows, as every new program benefits from the same structural efficiency, and a more accessible manufacturing profile supports a broader range of indications.
6.
Broad platform intellectual property. We have built a multi-layered intellectual property estate designed to protect cityRNA and establish what we believe is a new generation of RNAi platform intellectual property. The estate combines internally created, City Therapeutics-owned intellectual property with exclusively in-licensed foundational intellectual property from OSIF capturing the originating science that “cleavage-inducing tiny” RNAs can activate Argonaute-mediated target cleavage through mechanisms distinct from longer canonical siRNAs. Building on that foundation, our internal research has developed broad intellectual property spanning cityRNA molecules’ novel structural parameters, chemical modifications, AGO3 loading rules, tissue-specific delivery learnings, design rules and proprietary therapeutic trigger sequences. Together these are expected to create a robust intellectual property perimeter around the City Platform that strengthens and broadens with each scientific and clinical milestone we achieve.
AI/ML Based Molecular Design of Triggers
Trigger design has historically been time-consuming and driven by trial-and-error. We are changing that with a proprietary machine-learning capability designed to predict trigger potency in silico. From our
founding, we invested heavily in our data and have amassed a large, proprietary dataset of RNAi trigger measurements spanning a wide range of hepatic and extrahepatic targets. We use this dataset to train complementary models that identify the design rules driving activity and predict which novel candidates are most likely to succeed, letting us focus laboratory testing on our highest-potential trigger designs. This automated framework enables us to complete dozens of modeling experiments in days rather than the months such work previously required.
We believe this dataset and AI/ML capability is a valuable asset, and we will continue to build and invest in this important capability. Our initial modeling results already outperform the best published tools for predicting siRNA activity, and we are applying the same approach to cityRNAs, building a comprehensive sequence and chemistry activity dataset to identify the best cityRNA candidates. These models now drive a continuous design-build-test cycle in which each round of laboratory testing feeds the next round of predictions, so the capability improves with every experiment, compressing design timelines, expanding our proprietary RNAi trigger estate, and widening our lead in trigger engineering.
Novel Delivery Conjugates
Even the most potent RNAi trigger has little therapeutic value unless it can be delivered to the appropriate cells. Accordingly, our conjugate chemistry is designed to establish tissue selectivity and maximize productive intracellular delivery. For more than two decades, delivery, not target biology, has defined the therapeutic boundaries of RNAi. Whereas the liver is readily accessible, extrahepatic tissues present formidable extracellular, endothelial, and cellular barriers that demand tailored delivery strategies. Our approach uses established and proprietary ligands, such as GalNAc for hepatic delivery and LRP1 ligands for selected extrahepatic tissues, enabling efficient uptake and intracellular trafficking in the intended cell populations.
LRP1: reaching the eye, CNS, and adipose tissue. We have developed proprietary ligands for LRP1, the low-density lipoprotein receptor-related protein 1, a receptor we believe is particularly well suited to delivering RNAi triggers to ocular, CNS, and adipose tissues. LRP1 is highly expressed across multiple relevant cell populations, including neurons, astrocytes, oligodendrocyte precursor cells, and microglia; it undergoes rapid endocytosis and recycling, giving it high transport capacity; it is not known to trigger a signaling cascade upon ligand binding; and it is highly conserved across species, supporting translation from preclinical models to humans.
In experimental studies, LRP1 conjugates achieved broad retinal distribution and substantially deeper target knockdown than an unconjugated siRNA trigger following intravitreal administration, and drove robust gene silencing across multiple brain regions and in adipose tissue following intrathecal and systemic dosing, respectively (as shown in Figures 16 and 17). Together, these results support the potential for LRP1 as a high-capacity delivery system capable of enabling meaningful RNAi activity in tissues that have historically been difficult to access.
Figure 16. LRP1 ligand enabled broad retinal distribution and enhanced target mRNA knockdown compared to siRNA alone.
Figure 17. LRP1 ligand — conjugates drove robust gene silencing across multiple extrahepatic tissues, including brain and adipose.
Expanding the conjugate toolkit. Beyond GalNAc and LRP1, we are actively developing additional novel delivery technologies, including antibodies and peptides, to extend our City Platform into other tissues.
Target Discovery and Selection
Because RNAi can silence almost any gene, the choice of target is as consequential as the molecule that silences it, and we begin every program with biology supported by human genetic evidence. We have established a genetics capability that combines statistical genetic approaches with access to comprehensive human genetic data linked to electronic medical records of over 1 million participants to build confidence around any target that we nominate. Using this capability, we can perform analyses on any clinical or physical measurement to build genetic support for our targets, including evaluating target biology, assessing potential tolerability profiles, and informing our therapeutic hypotheses.
We have used this genetics-first discipline to design our pipeline: FXI, RBP4, and TFR2, are each genetically and/or clinically supported targets whose human biology pointed to potential tolerability and clinical activity. Paired with our RNAi trigger engineering and delivery conjugates, target discovery is designed to ensure that the therapeutics we build are aimed at biology supported by human genetic data.
Summary of the City Platform
The three capabilities of the City Platform reinforce one another: a genetically supported target, engineered into a potent and specific RNAi trigger molecule, delivered by a conjugate tuned to the tissue. Our goal is to build high potency and specificity into every development candidate we design through RNAi trigger engineering built upon our deep RNAi therapeutics expertise, our proprietary cityRNA capability, and AI/ML driven molecular design. We believe our City Platform is a durable competitive advantage that supports our goal of filing one to two new IND or IND-equivalent applications per year.
Licensing Agreements, Partnerships, Collaborations
Bausch + Lomb Research Collaboration and License Agreement
On July 8, 2024, we entered into a Research Collaboration and License Agreement (the “B+L Agreement”) with Bausch + Lomb. Pursuant to the B+L Agreement, we (i) agreed to undertake a research program to discover a certain licensed product intended to enable Bausch + Lomb to develop and subsequently commercialize such licensed product in the field of ocular disease and (ii) granted
Bausch + Lomb an option to obtain an exclusive, worldwide license to certain of our intellectual property to develop, manufacture and commercialize a licensed product in the field of ocular disease.
Under the terms of the B+L Agreement, the parties agreed to collaborate pursuant to a research plan under which we will use certain commercially reasonable efforts to perform specified research activities overseen by a joint research committee.
Bausch + Lomb has an option, exercisable for a specified period, to select one or more development candidates for further development and commercialization. Upon exercise of such option and payment of a certain development candidate selection fee, we agreed to grant Bausch + Lomb an exclusive, worldwide, royalty-bearing sublicensable license under the licensed technology to exploit a licensed product in the field of ocular disease. Following such election, Bausch + Lomb will have, at its expense, sole responsibility for development, manufacturing, regulatory, and commercialization activities related to the licensed product, subject to the use of certain commercially reasonable efforts.
During the term of the B+L Agreement, subject to specified exceptions, we are restricted from researching, developing, manufacturing or commercializing competing products directed to the collaboration target and, for a limited period, certain related targets and retinal disease programs.
Pursuant to the B+L Agreement, we received a non-refundable, non-creditable upfront payment of $5.0 million from Bausch + Lomb in July 2024. Simultaneously upon execution of the B+L Agreement, we issued a convertible note to Bausch + Lomb in an aggregate amount of $10.0 million, bearing interest at 8.0% per annum and maturing on July 8, 2027, unless earlier converted, which subsequently converted into shares of our Series B-1 convertible preferred stock.
We are also eligible to receive (i) a certain development candidate selection fee and certain other development, regulatory, commercial and sales milestones, collectively up to an aggregate of $485.0 million, and (ii) tiered royalties ranging from mid-single digits to low teens on annual net sales of licensed products, subject to customary reductions and an overall royalty floor. Upon expiration of the applicable royalty term, the license becomes fully paid-up, perpetual, irrevocable, and royalty-free.
If the agreement expires or terminates following specified stages of the collaboration and we subsequently commercialize a licensed molecule generated under the research program, we are required to pay Bausch + Lomb a low-single digit royalty on future net sales of products containing such licensed molecule for a fixed royalty term following first commercial sale.
Unless earlier terminated, the B+L Agreement will remain in force until the earlier of (i) the occurrence of any development candidate selection termination condition, or (ii) the expiration of the last-to-expire royalty term for all licensed products. Bausch + Lomb may terminate for convenience, in its entirety or on a country-by-country basis, upon prior written notice. Either party may terminate for an uncured material breach, subject to a specified cure period.
Biogen Research Collaboration and License Agreement
On May 23, 2025, we entered into a Research Collaboration and License Agreement (the “Biogen Agreement”) with Biogen. Pursuant to the Biogen Agreement, we agreed to collaborate with Biogen to undertake a research plan to discover and develop certain RNAi triggers that inhibit the expression of a target protein and conjugates of such RNAi triggers.
During the research term, each party agreed to grant the other a non-exclusive, royalty-free, worldwide license under certain research technology solely to perform research activities under an agreed research plan. We also granted Biogen an exclusive, worldwide, royalty-bearing license to certain of our intellectual property, with the right to sublicense through multiple tiers, to develop, manufacture and commercialize licensed compounds and licensed products in the field in the territory. However, unless and until Biogen elects one or more development candidates for further development and commercialization, Biogen is restricted from clinically developing or commercializing licensed compounds or licensed products. Following a development candidate selection, Biogen has sole responsibility, at its expense, for development, manufacturing, regulatory, medical affairs, and commercialization activities related to such selected development candidate.
Under the terms of the Biogen Agreement, the parties agreed to conduct research pursuant to a research plan overseen by a joint research committee and joint research team, with Biogen holding final decision-making authority, subject to specified limitations.
We also granted Biogen an option, exercisable during a specified period and upon specified terms, to designate one additional target, subject to execution of a separate collaboration and license agreement on substantially the same terms and an additional payment of a specified option exercise fee.
Pursuant to the terms of the Biogen Agreement, we received a non-refundable, non-creditable upfront payment of $16.0 million from Biogen in June 2025. We are eligible to receive an additional non-refundable upfront payment upon the satisfaction of certain conditions relating to our upstream license with OSIF. We remain responsible for payments due under our upstream license agreements. Simultaneously with the execution of the Biogen Agreement, we issued a convertible note to Biogen in an aggregate principal amount of $30.0 million, bearing interest at 8.0% per annum and maturing on July 1, 2027.
We are also eligible to receive up to an aggregate of approximately $1.0 billion in potential research, development, commercial, and sales milestones, along with tiered royalties ranging from high-single digits to low teens on annual net sales of licensed products, subject to customary reductions and an overall royalty floor. Upon expiration of the applicable royalty term, the license becomes fully paid-up, perpetual, irrevocable and royalty-free.
Unless earlier terminated, the Biogen Agreement will continue on a licensed product-by-licensed product and country-by-country basis, until the latest of: (i) the date on which there is no royalty-bearing patent claim with respect to such licensed product in such country, (ii) the expiration of all regulatory exclusivity for such licensed product in such country, and (iii) a specified number of years after the first commercial sale of such licensed product in such country.
Either party may terminate the agreement if Biogen has not made at least one RNAi trigger candidate selection or the development candidate selection by the applicable outside date. Biogen may terminate the agreement for convenience, in its entirety or on a country-by-country and licensed product-by-licensed product basis, upon prior written notice. Either party may also terminate for an uncured material breach, subject to a specified cure period, which may be extended upon submission of a reasonable cure plan.
OSIF Exclusive License Agreement
On October 30, 2023, we entered into the OSIF License Agreement with OSIF. Under the OSIF License Agreement, we obtained an exclusive, royalty-bearing license to certain patent rights relating to the originating technology underlying cityRNAs and certain other aspects of our technology platform to make, have made, use, sell, and import certain licensed products, subject to certain terms and restrictions therein.
Upon execution of the OSIF License Agreement, we issued equity securities representing 5% of our fully diluted capitalization as of the effective date of the agreement and agreed to issue additional equity securities to OSIF from time to time as necessary to maintain OSIF’s pro rata ownership percentage until we have received a specified amount of equity financing proceeds. The OSIF License Agreement has been subsequently amended to add additional patent rights and technology disclosures to the licensed intellectual property, and we reimbursed OSIF for patent prosecution costs associated with such additional patent rights.
Pursuant to the terms of the OSIF License Agreement, we are required to pay OSIF tiered low-single digit running royalties on annual net sales of each licensed product, subject to limited offsets for third-party royalty obligations. Following the first commercial sale of a licensed product in a major market, we are also required to pay OSIF a specified minimum annual royalty amount. In addition, we are required to pay OSIF a total of up to $4.2 million for each licensed product upon achievement of specified clinical development, regulatory approval, and commercial sales milestones.
The OSIF License Agreement also requires us to pay OSIF tiered percentages of certain non-royalty sublicense consideration, ranging from a low-twenties to mid-twenties percentage at earlier stages of development and a low-tens to mid-teens percentage at later stages of development, with the applicable percentage depending on the stage of development at which the sublicense is granted. In addition, we are responsible for certain patent costs incurred after the effective date of the agreement and annual maintenance fees.
Unless earlier terminated, the OSIF License Agreement continues until the expiration of the last to expire of the patent rights. At our option, we may terminate upon prior written notice. OSIF may terminate the agreement at its option immediately upon delivery of prior written notice if any specified events occur, including: (i) our failure to make any payment when due; (ii) breach of any non-payment provision; or (iii) if we or our affiliates or sublicensees initiate any proceeding to challenge the validity, enforceability, ownership, or scope of the patent rights, in each case pursuant to specified cure periods.
Intellectual Property
We view intellectual property as a core strategic asset and an important component of our strategy to lead the next generation of RNAi therapeutics. We have assembled, and continue to expand, a broad and multi-layered intellectual property portfolio covering our City Platform, which includes: our RNAi trigger toolkit (e.g., improved canonical siRNAs and our proprietary cityRNAs); delivery technologies; and RNAi chemistry. We have also assembled and continue to build a robust intellectual property portfolio covering our RNAi therapeutic candidates, and related methods of use. This portfolio is anchored in innovations generated through our internal research and development activities, including discoveries made by our scientists as they advance our City Platform and product candidates, together with intellectual property exclusively in-licensed from OSIF and other third-party rights that we may identify as strategically important to our business.
Our intellectual property strategy is deliberate, iterative, and designed to evolve as our City Platform and pipeline mature. We seek to capture and protect innovation generated within City across all stages of discovery and development, including improvements in RNAi trigger design, RNAi chemistry, extrahepatic delivery, RNAi therapeutic candidates, manufacturing processes, and other know-how that may be important to our current and future programs. At the same time, we actively monitor the external intellectual property landscape to identify third-party technologies, patent rights, know-how, and other assets that may be useful to complement, strengthen, or extend our proprietary position. Where appropriate, we may pursue licenses, collaborations, acquisitions, or other arrangements to access such third-party intellectual property.
Rather than relying on a single category of protection, we seek to build multiple, reinforcing layers of intellectual property around our City Platform and product candidates. These layers may include patent rights directed to RNAi trigger structures and designs; chemical modifications and motifs intended to improve potency, selectivity, durability, and tolerability; delivery ligands and conjugates designed for extrahepatic tissues; RNAi therapeutic compositions of matter and methods of treatment; manufacturing processes; and proprietary know-how and trade secrets arising from our internal research and development activities. We believe this layered approach is important to protecting the breadth of innovations underlying our City Platform and our current and future RNAi product candidates.
Our objective is to continue to expand, strengthen, and maintain our intellectual property portfolio as we advance our product candidates, further develop our City Platform, and generate additional data and technical insights. We intend to pursue patent protection in the United States and selected jurisdictions outside the United States where we believe protection is commercially important, and we may seek additional patent protection as we identify new platform innovations, including delivery technologies, chemistry motifs, RNAi trigger designs, and RNAi therapeutic candidates. We also rely, and expect to continue to rely, on trade secrets, confidential know-how, regulatory exclusivities, patent term extensions, and other available forms of intellectual property and regulatory protection to support our proprietary position.
As of September 17, 2026, our portfolio is comprised of 19 patent families, including at least 12 pending U.S. patent applications, four pending foreign patent applications in Europe and Hong Kong
and eight Patent Cooperation Treaty (“PCT”) applications that have not yet entered national phase. Any patents issuing from or claiming priority to these applications, if granted, are expected to expire between 2041 and 2047, excluding any patent term adjustment or extension and assuming payment of all required maintenance, renewal, and annuity fees.
In addition to patent rights, we rely on trade secrets and proprietary know-how relating to our City Platform, candidate-selection processes, manufacturing methods, screening approaches and other aspects of our research and development operations. We expect trade secrets and know-how to remain important.
City Platform Intellectual Property
Our City Platform-related intellectual property spans three principal categories: RNAi triggers, including improved canonical siRNA and cityRNA triggers; chemistry and related structural optimizations; and delivery molecules, including ligands designed to deliver to the eye, CNS and adipose tissue. Our platform portfolio includes owned and exclusively in-licensed patent rights and know-how intended to protect compositions, methods of design, methods of use and other innovations relevant to the next generation RNAi triggers and extrahepatic delivery molecules.
Our RNAi trigger-related portfolio is intended to protect both next generation canonical siRNAs and cityRNAs. Our chemistry-related intellectual property is intended to protect proprietary motifs and other design features relevant to City-engineered RNAi trigger and intended to enhance potency, selectivity, durability or tissue reach. Our delivery-related intellectual property is intended to protect ligand compositions and related methods directed to extrahepatic delivery, including technologies relevant to the delivery to the liver, eye, CNS and adipose tissue. Taken together, we believe these rights support a layered protection strategy around the modular architecture of our City Platform.
The City Platform-related intellectual property relates broadly to our existing RNAi therapeutic candidates and those we may develop in the future and the indications we target or may target in the future. We in-licensed patent rights related to cityRNAs from OSIF. As of September 17, 2026, we in-licensed four non-provisional U.S. patent applications, two pending PCT patent applications, and four foreign patent applications in Europe and Hong Kong. We expect patents issuing from or claiming priority to these pending applications, if any, to expire between 2041 and 2046, excluding any patent term adjustments or extensions.
We have also filed one City owned pending U.S. provisional application and one City owned PCT International Patent Application directed to compositions and methods relating to City’s improvements to cityRNAs and two City owned U.S. provisional applications directed to compositions and methods directed to City’s improvements to canonical siRNAs. We expect patents issuing from or claiming priority to these pending applications, if any, to expire between 2046 and 2047, excluding any patent term adjustments or extensions.
In addition, we own one PCT patent application and one pending U.S. provisional application directed to LRP1 targeting ligands for extrahepatic delivery. We expect patents issuing from or claiming priority to these pending applications, if any, to expire in 2046, excluding any patent term adjustments, or extensions.
Therapeutic/Candidate Intellectual Property
Our RNAi candidate-related patent filings are intended to protect RNAi compositions, formulations, methods of use, and related inventions directed to our current and future therapeutic programs.
CITY-FXI
For CITY-FXI, our filings are intended to protect siRNA candidates directed to reducing expression of FXI in thrombotic disease, including relevant compositions and therapeutic uses. As of September 17, 2026, we own one pending U.S. provisional patent application and one PCT application related to
siRNA compositions of matter and methods of treatment. We expect patents issuing from these pending patent applications, if any, to expire between 2045 and 2047, excluding any patent term adjustments or extensions.
CITY-RBP4
For CITY-RBP4, our filings are intended to protect siRNA candidates directed to RBP4 inhibition for the treatment of ocular disease, including Stargardt disease and other retinal indications. As of September 17, 2026, we owned one PCT application and one U.S. provisional patent application related to siRNA compositions of matter and methods of treating ocular disease. We expect patents issuing from or claiming priority to these pending patent applications, if any, to expire between 2046 and 2047, excluding any patent term adjustments or extensions.
CITY-TFR2
For CITY-TFR2, our filings are intended to protect siRNA candidates directed to reducing expression of TFR2 for anemia indications, including myelofibrosis-associated anemia and other chronic inflammatory conditions we may pursue. As of September 17, 2026, we own one PCT application related to siRNA compositions of matter and methods of treating anemia of chronic disease. We expect patents issuing from or claiming priority to this application, if any, to expire in 2046, excluding any patent term adjustments or extensions.
Other Disease Areas
In addition to our disease programs listed above, we also have patent applications relating to novel RNAi therapeutic candidates and their use for treating additional disorders that would benefit from inhibition. As of September 17, 2026, we own patent applications directed to RNAi compositions and methods of treatment for neurological and ocular disorders. We expect patents issuing from or claiming priority to these pending applications, if any, to expire between 2046 and 2047 excluding any patent term adjustments or extensions.
We also file, and expect to continue to file, patent applications directed to discovery-stage programs and other disease areas that may benefit from our platform technologies. Our objective is to maintain a portfolio that protects both current clinical-stage candidates and future programs arising from our City Platform.
Manufacturing and Supply
We do not own or operate, and currently have no plans to establish, any manufacturing facilities. We have engaged, and expect to continue to rely on, well-established third-party contract development and manufacturing organizations (“CDMOs”) to supply our product candidates for use in our preclinical studies and clinical trials. Because we rely on CDMOs, we employ personnel with extensive technical, manufacturing, analytical, and quality experience to oversee contract manufacturing and testing activities, and to compile manufacturing and quality information for our regulatory submissions. We believe our CDMOs have the scale, systems, and experience to supply our currently planned clinical trials.
Additionally, we intend to rely on third-party CDMOs for later-stage development and commercial manufacturing, if our product candidates receive marketing approval. As our lead product candidates advance through clinical development, we expect to enter into longer-term commercial supply agreements to fulfill and secure our production needs. While the drug substances used in our product candidates are manufactured by more than one supplier, the number of manufacturers is limited. In the event it is necessary or advisable to acquire supplies from an alternative supplier, we might not be able to obtain them on commercially reasonable terms, if at all. It could also require significant time and expense to redesign our manufacturing processes to work with another company. If we need to change manufacturers during the clinical or development stage for product candidates or after commercialization for our product candidates, if approved, the FDA, European Medicines Agency (“EMA”) and corresponding foreign regulatory agencies must approve these new manufacturers in advance, which will involve testing
and additional inspections to ensure compliance with FDA regulations and standards and may require significant lead times and delay. Reliance on third-party CDMOs may expose us to different risks than if we were to manufacture and develop product candidates ourselves. Should any of these manufacturers become unavailable to us for any reason, we believe that there are a number of potential replacements, although we may incur some delay in identifying and qualifying such replacements.
To adequately meet our projected commercial manufacturing needs, our CDMOs will need to scale-up production, or we will need to secure additional suppliers. Processes for producing drug substances and drug product for commercial supply are currently being developed, with the goal of achieving reliable, reproducible, and cost-effective production. We believe the drug substance and drug product processes for our current product candidates can be appropriately scaled.
Commercialization
Given our stage of development, we have not yet established a commercial organization or distribution capabilities. We plan to independently commercialize our products, if approved, in the United States and other regions where we determine it makes commercial sense to do so. At the appropriate time, we will recruit a sales force and a medical affairs team and take other steps to establish the necessary commercial infrastructure. However, as product candidates advance through our pipeline, our plans may change.
Competition
The biopharmaceutical industry is characterized by rapidly advancing technologies, intense competition, and a strong emphasis on proprietary and novel products and product candidates. We face a broad spectrum of current and potential competitors, ranging from very large, global pharmaceutical companies with significant resources to other biotechnology companies with resources and expertise comparable to our own. We believe that for most or all of our drug development programs, there will be one or more competing programs in other companies. In many cases, the companies with competing programs will have access to greater resources and expertise than we do and may be more advanced.
As RNAi therapeutics expand beyond rare diseases to other more common conditions, an increasing number of other companies are investing in and developing RNAi therapeutic products, as well as potentially new approaches that may result in more rapid development of RNAi therapeutics or more effective technologies for RNAi drug development or delivery.
Companies that have developed, or are developing, siRNA therapeutics include ADARx Pharmaceuticals, Inc., Alnylam, Amgen Inc., Argo Biopharmaceutical Co., Ltd., Aro Biotherapeutics Co., Arrowhead Pharmaceuticals, Inc., AstraZeneca plc, Dyne Therapeutics, Inc., Eli Lilly and Company, GlaxoSmithKline plc, Ionis Pharmaceuticals, Inc., Novartis AG, Novo Nordisk A/S, SanegeneBio Inc., Sarepta Therapeutics, Inc., Silence Therapeutics plc, and Sirius Therapeutics, Inc.
Our competitors have developed, are developing or may develop product candidates and products competitive with CITY-FXI, CITY-RBP4, and CITY-TFR2 and our other programs and product candidates. CITY-FXI, CITY-RBP4, and CITY-TFR2 and any future product candidates that we successfully develop and commercialize will compete with existing therapies and new therapies that may become available in the future.
Our lead product candidate, CITY-FXI, is being developed for thromboembolic diseases through silencing of coagulation FXI. Multiple therapeutic approaches targeting FXI are currently in development, including small molecules, monoclonal antibodies, and RNAi therapeutics. Companies active in this field include ADARx Pharmaceuticals, Inc., Bayer AG, Bristol-Myers Squibb Company, CRISPR Therapeutics AG (in partnership with Sirius Therapeutics, Inc.), Johnson & Johnson, Novartis AG, Regeneron Pharmaceuticals, Inc., and others.
CITY-RBP4 is being developed for Stargardt disease, for which current treatment options are limited. Numerous companies are developing potential therapies for Stargardt disease using a variety
of approaches, including visual cycle modulation, gene therapy, gene editing, and RNA-based therapeutics. Companies active in this area include Alkeus Pharmaceuticals, Inc., Belite Bio, Inc. and others.
CITY-TFR2 is being developed for anemia of chronic disease. Therapeutic approaches under development or in clinical use include hepcidin-targeted agents, including a monoclonal antibody from Disc Medicine, Inc., erythropoiesis-stimulating agents, hypoxia-inducible factor prolyl hydroxylase inhibitors, iron replacement therapies, and other approaches intended to improve iron availability or erythropoiesis.
Our competitors include larger and better-funded global pharmaceutical and biotechnology companies. Smaller or early-stage companies may also prove to be significant competitors, particularly through collaborative arrangements with large and established companies. Moreover, we may also compete with universities, governmental agencies, and other public and private research institutions that may be active in research in our target indications and could be in direct competition with us. We also compete with these organizations to recruit management, scientists, and clinical development personnel, and our inability to compete successfully could negatively affect our level of expertise and our ability to execute our business plan. We will also face competition in establishing clinical trial sites and enrolling subjects for clinical trials and in identifying and in-licensing intellectual property related to new product candidates, as well as entering into partnerships, collaborations, and license arrangements.
If our product candidates do not offer advantages over available products, we may not be able to successfully compete against current and future competitors. The key factors affecting the success of our products, if approved, are likely to be their potential efficacy, safety, convenience, and availability of reimbursement.
Government Regulation
Government authorities in the United States, at the federal, state, and local level, and in other countries and jurisdictions, including the European Union (“EU”), extensively regulate, among other things, the research, development, testing, manufacturing, quality control, safety, effectiveness, approval, packaging, storage, recordkeeping, labeling, advertising, promotion, distribution, marketing, post-approval monitoring and reporting, and import and export of pharmaceutical products. The processes for obtaining regulatory approvals in the United States and in foreign countries and jurisdictions, along with subsequent compliance with applicable statutes and regulations, require the expenditure of substantial time and financial resources. Even after obtaining initial marketing approval, a product and its manufacturer remain subject to extensive, continuing regulatory requirements, including with respect to manufacturing, quality control, adverse event reporting, advertising and promotion and periodic inspections by regulatory authorities.
Review and Approval of Drugs in the United States
In the United States, the FDA regulates drugs under the U.S. Federal Food, Drug, and Cosmetic Act (“FDCA”), and its implementing regulations. The failure to comply with applicable U.S. requirements at any time during the product development process, approval process, or after approval may subject an applicant and/or sponsor to a variety of administrative or judicial sanctions, including refusal by the FDA to approve pending applications, withdrawal of an approval, imposition of a clinical hold, issuance of warning letters, FDA Form 483s, and other types of letters, product seizures, total or partial suspension of production or distribution, injunctions, fines, product recalls, and other post-market actions, refusals of government contracts, restitution, disgorgement of profits, or civil or criminal investigations, and penalties brought by the FDA and the U.S. Department of Justice or other governmental entities.
An applicant seeking approval to market and distribute a new drug product in the United States must typically undertake the following:
•
completion of nonclinical, or preclinical, laboratory tests, animal studies, and formulation studies in compliance with the FDA’s good laboratory practice (“GLP”) regulations;
•
submission to the FDA of an IND, which must take effect before human clinical trials may begin;
•
approval by an institutional review board (“IRB”), representing each clinical site before each clinical trial may be initiated at that site;
•
performance of adequate and well-controlled human clinical trials in accordance with good clinical practices (“GCPs”), to establish the safety and efficacy of the proposed drug product for each indication;
•
preparation and submission to the FDA of a New Drug Application (“NDA”), and payment of user fees;
•
review of the product by an FDA advisory committee, where appropriate or if applicable;
•
satisfactory completion of one or more FDA inspections of the manufacturing facility or facilities at which the product, or components thereof, are produced to assess compliance with current Good Manufacturing Practices (“cGMP”) requirements and to assure that the facilities, methods and controls are adequate to preserve the product’s identity, strength, quality, and purity;
•
satisfactory completion of FDA audits of clinical trial sites to assure compliance with GCPs and the integrity of the clinical data; and
•
FDA review and approval of the NDA.
Preclinical Studies
Before an applicant begins testing a compound in humans, the drug candidate enters the preclinical testing stage. Preclinical studies include, among other things, laboratory evaluation of the purity and stability of the manufactured drug substance or active pharmaceutical ingredient (“API”), and the formulated drug or drug product, as well as in vitro and animal studies to assess the potential safety and activity of the drug for initial testing in humans and to establish a rationale for therapeutic use. The conduct of certain preclinical studies is subject to federal regulations and requirements, including GLP regulations. Some long-term preclinical testing, such as animal tests of reproductive adverse events (“AEs”), and carcinogenicity, may continue after the IND is submitted.
The IND and IRB Processes
An IND is an exemption from the FDCA that allows an investigational drug to be shipped in interstate commerce for use in a clinical trial and a request for FDA authorization to administer such investigational drug to humans. Such authorization must be secured prior to interstate shipment and administration of the investigational drug. In an IND, applicants must submit a protocol for each clinical trial and any subsequent protocol amendments. In addition, the results of the preclinical tests, manufacturing information, analytical data, any available clinical data or literature and plans for clinical trials, among other things, are submitted to the FDA as part of an IND. An IND automatically becomes effective 30 days after receipt by the FDA, unless before that time, the FDA raises concerns or questions related to one or more proposed clinical trials and places the trial on clinical hold. The FDA also may impose a clinical hold or partial clinical hold after commencement of a clinical trial under an IND. A clinical hold is an order issued by the FDA to the sponsor to delay a proposed clinical investigation or to suspend an ongoing investigation. A partial clinical hold is a delay or suspension of only part of the clinical work requested under the IND. No more than 30 days after imposition of a clinical hold or partial clinical hold, the FDA will provide the sponsor a written explanation of the basis for the hold. Following issuance of a clinical hold or partial clinical hold, an investigation (or full investigation in the case of a partial clinical hold) may only resume after the FDA has notified the sponsor that the investigation may proceed. The FDA will base that determination on information provided by the sponsor correcting the deficiencies previously cited or otherwise satisfying the FDA that the investigation can proceed.
A sponsor may choose, but is not required, to conduct a foreign clinical trial under an IND. When a foreign clinical trial is conducted under an IND, all FDA IND requirements must be met unless waived.
When the foreign clinical trial is not conducted under an IND, the sponsor must ensure that the study is conducted in accordance with GCP, among other things, including review and approval by an independent ethics committee and obtaining informed consent from subjects. The GCP requirements are intended to help ensure the protection of human subjects enrolled in non-IND foreign clinical trials, as well as the quality and integrity of the resulting data. FDA must also be able to validate the data from the study through an on-site inspection if necessary.
In addition to the foregoing IND requirements, an IRB representing each institution participating in the clinical trial must review and approve the plan for any clinical trial before it commences at that institution, and the IRB must conduct continuing review of the study at least annually. The IRB must review and approve, among other things, the study protocol and informed consent information to be provided to study subjects. An IRB must operate in compliance with FDA regulations. An IRB can suspend or terminate approval of a clinical trial at its institution, or an institution it represents, if the clinical trial is not being conducted in accordance with the IRB’s requirements or if the product candidate has been associated with unexpected serious harm to patients.
Additionally, some trials are overseen by an independent group of qualified experts organized by the trial sponsor, known as a data safety monitoring board or committee. This group provides advice to the sponsor as to whether or not a trial may move forward at designated check points based on pre-specified criteria and access that only the group maintains to available data from the study. The FDA or the sponsor may suspend or terminate a clinical trial at any time on various grounds, including a finding that the research subjects are being exposed to an unacceptable health risk.
Information about certain clinical trials must be submitted within specific timeframes to the National Institutes of Health for public dissemination on its ClinicalTrials.gov website. Sponsors also must disclose certain results of these clinical trials, although disclosure of results may be delayed until after the new product or new indication has been approved by the FDA. Competitors may use this publicly available information to gain knowledge regarding the progress of development programs, as well as clinical trial design. Failure to timely register a covered clinical study or to submit study results as provided for in the law can give rise to public notifications of noncompliance, civil monetary penalties, and also prevent the non-compliant party from receiving future grant funds from the federal government.
Human Clinical Trials in Support of an NDA
Clinical trials involve the administration of the investigational product to human subjects under the supervision of qualified investigators in accordance with GCP requirements, which include, among other things, the requirement that all research subjects, or their legal representative, provide their informed consent in writing before their participation in any clinical trial. Clinical trials are conducted under written study protocols detailing, among other things, the inclusion and exclusion criteria, the objectives of the study, the parameters to be used in monitoring safety and the effectiveness criteria to be evaluated.
Human clinical trials are typically conducted in three sequential phases, which may overlap or be combined:
•
Phase 1. The product candidate is initially introduced into healthy human subjects or in patients with the targeted disease or condition in certain indications such as cancer to test for safety, dosage tolerance, absorption, metabolism, distribution, excretion, and, if possible, to gain an early indication of its effectiveness.
•
Phase 2. The product candidate is administered to a limited patient population with the specified disease or condition to identify possible AEs and safety risks, to preliminarily evaluate the efficacy of the product for specific targeted diseases or conditions, and to determine dosage tolerance and optimal dosage.
•
Phase 3. The product candidate is administered to an expanded patient population, generally at geographically dispersed clinical trial sites, in well-controlled clinical trials with the intent to generate enough data to evaluate the efficacy and safety of the product for approval, to establish the overall risk-benefit profile of the product candidate and to provide adequate information for the labeling of the product.
Post-approval studies, sometimes referred to as Phase 4 studies, may be conducted after initial regulatory approval. These studies are used to gain additional experience from the treatment of patients in the intended therapeutic indication. In certain instances, the FDA may mandate the performance of Phase 4 clinical trials as a condition of approval of an NDA.
While the IND is active, progress reports detailing the results of the clinical trials, among other information, must be submitted at least annually to the FDA. In addition, within 15 calendar days after the sponsor determines that the information qualifies for reporting, IND safety reports must be submitted to the FDA for any of the following: serious and unexpected suspected adverse reactions; findings from other studies or animal or in vitro testing that suggest a significant risk in humans exposed to the drug; and any clinically important increase in the case of a serious suspected adverse reaction over that listed in the protocol or investigator brochure. The sponsor also must notify the FDA of any unexpected fatal or life-threatening suspected adverse reaction within seven calendar days after the sponsor’s initial receipt of the information.
Concurrent with clinical trials, companies often complete additional animal studies and must also develop additional information about the chemistry and physical characteristics of the drug as well as finalize a process for manufacturing the product in commercial quantities in accordance with cGMP requirements. The manufacturing process must be capable of consistently producing quality batches of the drug candidate and, among other things, the applicant must develop methods for testing the identity, strength, quality, purity, and potency of the final drug. Additionally, appropriate packaging must be selected and tested and stability studies must be conducted to demonstrate that the drug candidate does not undergo unacceptable deterioration over its shelf life.
Review of an NDA by the FDA
Assuming successful completion of required clinical testing and other requirements, the results of the preclinical studies and clinical trials, together with detailed information relating to the product’s chemistry, manufacture, controls, and proposed labeling, among other things, are submitted to the FDA as part of an NDA requesting approval to market the drug product for one or more indications. Under federal law, the submission of most NDAs is additionally subject to a significant application user fee as well as annual prescription drug product program fees. These fees are typically increased annually. Certain exceptions and waivers are available for some of these fees.
In addition, the Pediatric Research Equity Act (“PREA”) requires a sponsor to conduct pediatric clinical trials for most drugs, for a new active ingredient, new indication, new dosage form, new dosing regimen or new route of administration, and requires a sponsor to submit an initial Pediatric Study Plan, (“PSP”), within 60 days of an end-of-Phase 2 meeting or, if there is no such meeting, as early as practicable before the initiation of the Phase 3 or Phase 2/3 study. The initial PSP must include an outline of the pediatric study or studies that the sponsor plans to conduct, including study objectives and design, age groups, relevant endpoints and statistical approach, or a justification for not including such detailed information, and any request for a deferral of pediatric assessments or a full or partial waiver of the requirement to provide data from pediatric studies along with supporting information. The FDA and the sponsor must reach an agreement on the PSP. Under PREA, NDAs, and certain supplements must contain a pediatric assessment unless the sponsor has received a deferral or waiver. The required assessment must evaluate the safety and effectiveness of the product for the claimed indications in all relevant pediatric subpopulations and support dosing and administration for each pediatric subpopulation for which the product is deemed safe and effective. The sponsor or FDA may request a deferral of pediatric clinical trials for some or all of the pediatric subpopulations. A deferral may be granted for several reasons, including a finding that the product candidate is ready for approval for use in adults before pediatric clinical trials are complete or that additional safety or effectiveness data needs to be collected before any pediatric clinical trials begin. The FDA must send a non-compliance letter to any sponsor that fails to submit the required assessment, keep a deferral current or fails to submit a request for approval of a pediatric formulation.
The FDA conducts a preliminary review of an NDA within 60 days of its receipt, before accepting the NDA for filing, to determine whether the application is sufficiently complete to permit substantive review. The FDA may request additional information rather than accept an NDA for filing. In this event,
the application must be resubmitted with the additional information. The resubmitted application is also subject to review before the FDA accepts it for filing. Once the submission is accepted for filing, the FDA begins an in-depth substantive review. The FDA reviews an NDA to determine, among other things, whether a product is safe and effective for its intended use and whether its manufacturing is cGMP compliant to assure and preserve the product’s identity, strength, quality, and purity. Under the performance goals and policies implemented by the FDA under the Prescription Drug User Fee Act (“PDUFA”), FDA has agreed to specified performance goals in the review process of NDAs. Applications for drugs containing new molecular entities are meant to be reviewed within 10 months from the date of filing, and applications for “priority review” products containing new molecular entities are meant to be reviewed within six months of filing. The review process may be extended by the FDA for three additional months if FDA requests or the applicant otherwise provides additional information or clarification regarding information already provided in the submission within the last three months before the PDUFA goal date or is otherwise deemed a “major amendment” to the application.
During its review of an NDA, the FDA typically will inspect the facility or facilities where the product is or will be manufactured. These pre-approval inspections may cover all facilities associated with an NDA, including drug component manufacturing (such as APIs), finished drug product manufacturing, and control testing laboratories. The FDA will not approve an NDA unless it determines that the manufacturing processes and facilities are in compliance with cGMP requirements and adequate to assure consistent production of the product within required specifications. In addition, the FDA will typically inspect one or more clinical sites to assure compliance with GCP and the integrity of the clinical data submitted.
In addition, as a condition of approval, the FDA may require an applicant to develop a risk evaluation and mitigation strategy (“REMS”). REMS use risk minimization strategies beyond the professional labeling to ensure that the benefits of the product outweigh the potential risks. To determine whether a REMS is needed, the FDA will consider the size of the population likely to use the product, seriousness of the disease, expected benefit of the product, expected duration of treatment, seriousness of known or potential AEs, and whether the product is a new molecular entity. REMS can include medication guides, physician communication plans for healthcare professionals, and elements to assure safe use, or ETASU. ETASU may include, but are not limited to, special training or certification for prescribing or dispensing, dispensing only under certain circumstances, special monitoring, and the use of patient registries. The FDA may require a REMS before approval or post-approval if it becomes aware of a serious risk associated with use of the product.
The FDA is required to refer an application for a novel drug to an advisory committee or explain why such referral was not made. Typically, an advisory committee is a panel of independent experts, including clinicians and other scientific experts, that reviews, evaluates, and provides a recommendation as to whether the application should be approved and under what conditions. The FDA is not bound by the recommendations of an advisory committee, but it considers such recommendations carefully when making decisions.
Fast Track, Breakthrough Therapy, and Priority Review
The FDA has a number of programs intended to facilitate and expedite development and review of investigational drugs if they are intended to address an unmet medical need in the treatment of a serious or life-threatening disease or condition. Three of these programs are referred to as Fast Track Designation, Breakthrough Therapy Designation, and priority review designation.
First, the FDA may designate a product candidate for Fast Track review if it is intended, whether alone or in combination with one or more other products, for the treatment of a serious or life-threatening disease or condition, and it demonstrates the potential to address unmet medical needs for such disease or condition. For Fast Track product candidates, sponsors may have greater interactions with the FDA and the FDA may initiate review of sections of a Fast Track product candidate’s application before the application is complete. This rolling review may be available if the FDA determines, after preliminary evaluation of clinical data submitted by the sponsor, that a Fast Track product candidate may be effective. The sponsor must also provide, and the FDA must approve, a schedule for the submission of the remaining information and the sponsor must pay applicable user fees. However, the FDA’s
time period goal for reviewing a Fast Track application does not begin until the last section of the application is submitted. In addition, the Fast Track Designation may be withdrawn by the FDA if the FDA believes that the designation is no longer supported by data emerging in the clinical trial process.
Second, a product candidate may be designated as a Breakthrough Therapy if it is intended, either alone or in combination with one or more other drug products, to treat a serious or life-threatening disease or condition and preliminary clinical evidence indicates that the product candidate may demonstrate substantial improvement over existing therapies on one or more clinically significant endpoints, such as substantial treatment effects observed early in clinical development. Breakthrough Therapy Designation includes all of the benefits associated with Fast Track Designation, including the potential for rolling review of an NDA submission. In addition, the FDA may take certain actions with respect to Breakthrough Therapies, including holding meetings with the sponsor throughout the development process; providing timely advice to the product sponsor regarding development and approval; involving more senior staff in the review process; assigning a cross-disciplinary project lead for the review team; and taking other steps to design the clinical trials in an efficient manner.
Third, the FDA may designate an NDA review for a priority review if it is for a product candidate that treats a serious or life-threatening disease or condition and, if approved, would provide a significant improvement in safety or effectiveness compared to available therapies. The FDA determines, on a case-by-case basis, whether the product candidate may represent a significant improvement when compared with other available therapies. Significant improvement may be illustrated, among other things, by evidence of increased effectiveness in the treatment of a condition, elimination or substantial reduction of a treatment-limiting product reaction, documented enhancement of patient compliance that may lead to improvement in serious outcomes, and evidence of safety and effectiveness in a new subpopulation. A priority designation is intended to direct overall attention and resources to the evaluation of such applications, and to shorten the FDA’s goal for taking action on a marketing application from 10 months to six months.
Accelerated Approval Pathway
The FDA may grant accelerated approval to a product candidate intended to treat a serious or life-threatening condition that provides meaningful therapeutic advantage to patients over existing treatments based upon a determination that the product candidate has an effect on a surrogate endpoint that is reasonably likely to predict clinical benefit or on an intermediate clinical endpoint that can be measured earlier than an effect on irreversible morbidity or mortality (“IMM”), and that is reasonably likely to predict an effect on IMM or other clinical benefit, taking into account the severity, rarity, or prevalence of the condition and the availability or lack of alternative treatments. Products granted accelerated approval must meet the same statutory standards for safety and effectiveness as those granted traditional approval.
For the purposes of accelerated approval, a surrogate endpoint is a marker, such as a laboratory measurement, radiographic image, physical sign, or other measure that is thought to predict clinical benefit, but is not itself a measure of clinical benefit. Surrogate endpoints can often be measured more easily or more rapidly than clinical endpoints. An intermediate clinical endpoint is a measurement of a therapeutic effect that is considered reasonably likely to predict the clinical benefit of a product, such as an effect on IMM. The FDA has limited experience with accelerated approvals based on intermediate clinical endpoints, but has indicated that such endpoints generally may support accelerated approval where the therapeutic effect measured by the endpoint is not itself a clinical benefit and basis for traditional approval, if there is a basis for concluding that the therapeutic effect is reasonably likely to predict the ultimate clinical benefit of a product.
The accelerated approval pathway is often used in settings in which the course of a disease is long and an extended period of time is required to measure the intended clinical benefit of a product, even if the effect on the surrogate or intermediate clinical endpoint occurs rapidly.
The accelerated approval pathway is typically contingent on a sponsor’s agreement to conduct, in a diligent manner, additional post-approval confirmatory studies to verify and describe the product’s anticipated clinical benefit. As a result, a product candidate approved on this basis is subject to rigorous
post-marketing compliance requirements, including the completion of Phase 4 or other post-approval clinical trials to verify and describe the anticipated the effect on the clinical endpoint. Under the Food and Drug Omnibus Reform Act of 2022 (“FDORA”), the FDA is permitted to require, as appropriate, that such trials be underway prior to approval or within a specific time period after the date of approval for a product granted accelerated approval. Sponsors are also required to send updates to the FDA every 180 days on the status of such studies, including progress toward enrollment targets, and the FDA must post this information publicly. Under FDORA, the FDA has increased authority for expedited procedures to withdraw approval of a drug or indication approved under accelerated approval if, for example, the sponsor fails to conduct such studies in a timely manner and send the necessary updates to the FDA, or if a confirmatory trial fails to verify the predicted clinical benefit of the product. In addition, the FDA generally requires, unless otherwise informed by the agency, pre-approval of promotional materials for product candidates approved under accelerated regulations, which could adversely impact the timing of the commercial launch of the product.
Orphan Drug Designation
Under the Orphan Drug Act, the FDA may grant orphan designation to a drug intended to treat a rare disease or condition, which is a disease or condition that affects fewer than 200,000 individuals in the United States or, if it affects more than 200,000 individuals in the United States, there is no reasonable expectation that the cost of developing and making a drug product available in the United States for this type of disease or condition will be recovered from sales of the product. Orphan designation must be requested before submitting an NDA. After the FDA grants Orphan designation, the identity of the therapeutic agent and its potential orphan use are disclosed publicly by the FDA. Orphan designation does not convey any advantage in or shorten the duration of the regulatory review and approval process.
If a product that has orphan designation subsequently receives the first FDA approval for the disease or condition for which it has such designation, the product is entitled to orphan exclusivity, which means that the FDA may not approve any other applications to market the same therapeutic agent for the same approved use or indication for seven years, except in limited circumstances, such as a subsequent product’s showing of clinical superiority to the product with orphan exclusivity or inability of the original applicant to manufacture the product in sufficient quantities of the orphan drug to meet the needs of patients with the disease or condition for which the drug was designated. Orphan drug exclusivity does not prevent the FDA from approving a different drug for the same approved indication or use, or the same drug for a different indication or use. Among the other benefits of orphan drug designation are tax credits for certain research and a waiver of the NDA application user fee.
A designated orphan drug may not receive orphan drug exclusivity if it is approved for a use that is broader than the disease or condition for which it received orphan designation. In addition, orphan drug exclusive marketing rights in the United States may be lost if the FDA later determines that the request for designation was materially defective or, as noted above, if a second applicant demonstrates that its product is clinically superior to the approved product with orphan exclusivity within the relevant approved use or indication or the manufacturer of the approved product is unable to assure sufficient quantities of the product to meet the needs relating to the approved use or indication of patients with the relevant rare disease or condition.
Rare Pediatric Disease Designation and Priority Review Vouchers
Under the FDCA, the FDA incentivizes the development of drugs and biological products that meet the definition of a “rare pediatric disease,” defined to mean a serious or life-threatening disease in which the serious or life-threatening manifestations primarily affect individuals aged from birth to 18 years and the disease affects fewer than 200,000 individuals in the United States or affects more than 200,000 in the United States and for which there is no reasonable expectation that the cost of developing and making in the United States a drug or biological product for such disease or condition will be recovered from sales in the United States of such drug or biological product. The sponsor of a product candidate for a rare pediatric disease may be eligible for a voucher that can be used to obtain a priority review for a subsequent human drug or biological product application after the date of approval of the rare pediatric disease drug or biological product, referred to as a PRV. A sponsor may request rare pediatric
disease designation from the FDA prior to the submission of its NDA. A rare pediatric disease designation does not guarantee that a sponsor will receive a PRV upon approval of its NDA. Moreover, a sponsor who chooses not to submit a rare pediatric disease designation request may nonetheless receive a PRV upon approval of their marketing application if they request such a voucher in their original marketing application and meet all of the eligibility criteria. If a PRV is received, it may be sold or transferred an unlimited number of times. Congress has extended the PRV program through September 30, 2029. Under current law, after September 30, 2029, the FDA may not award any rare pediatric disease PRVs, although the FDA’s authority to do so could be extended by Congress in the future.
The FDA’s Decision on an NDA
On the basis of the FDA’s evaluation of the NDA and accompanying information, including the results of any inspections of the manufacturing facilities and select clinical trial sites, the FDA may issue an approval letter or a complete response letter. An approval letter authorizes commercial marketing of the product with specific prescribing information for specific indications. A complete response letter generally outlines the deficiencies in the submission and may require substantial additional testing or information in order for the FDA to reconsider the application. If a complete response letter is issued, the applicant may resubmit the NDA to address all of the deficiencies identified in the letter, withdraw the application, or request a hearing. If the applicant resubmits the NDA, the FDA will issue an approval letter only when the deficiencies have been addressed to the FDA’s satisfaction. The FDA has committed to reviewing such resubmissions in two or six months depending on the type of information included. Even with submission of this additional information, the FDA ultimately may decide that the application does not satisfy the regulatory criteria for approval. In September 2025, the FDA began publishing complete response letters soon after issuing them to the respective sponsors, breaking with long standing agency tradition of publishing complete response letters with approval documentation after the product is approved.
If the FDA approves a product, it may limit the approved indications for use for the product, require that contraindications, warnings, or precautions be included in the product labeling, require that post-approval studies, including Phase 4 clinical trials, be conducted to further assess the drug’s safety or effectiveness after approval, require testing and surveillance programs to monitor the product after commercialization, or impose other conditions, including distribution restrictions or other risk management mechanisms, including REMS, which can materially affect the potential market and profitability of the product. The FDA may prevent or limit further marketing of a product based on the results of post-market studies or surveillance programs.
Post-Approval Requirements
Drugs manufactured or distributed pursuant to FDA approvals are subject to pervasive and continuing regulation by the FDA, including, among other things, requirements relating to recordkeeping, periodic reporting, product sampling and distribution, advertising, and promotion, reporting of adverse experiences with the product and applicable product tracking and tracing requirements. After approval, many changes to the approved product, such as adding new indications or other labeling claims, are subject to prior FDA review and approval. There also are annual prescription drug product program fee requirements for certain marketed products.
In addition, drug manufacturers and other entities involved in the manufacture and distribution of approved drugs, and those supplying products, ingredients, and components of them are required to register their establishments with the FDA and state agencies, and are subject to periodic unannounced inspections by the FDA and these state agencies for compliance with cGMP requirements. Changes to the manufacturing process are strictly regulated and often require prior FDA approval before being implemented. FDA regulations also require investigation and correction of any deviations from cGMP and impose reporting and documentation requirements upon the NDA holder and any third-party manufacturers that the NDA holder may decide to use. Manufacturers and other parties involved in the drug supply chain for prescription drug products must also comply with product tracking and tracing requirements and notify the FDA of counterfeit, diverted, stolen, and intentionally adulterated products
or products that are otherwise unfit for distribution in the United States. Accordingly, manufacturers must continue to expend time, money, and effort in the area of production and quality control to maintain cGMP compliance.
Once an approval is granted, the FDA may withdraw the approval if compliance with regulatory requirements and standards is not maintained or if problems occur after the product reaches the market. Later discovery of previously unknown problems with a product, including AEs of unanticipated severity or frequency, or with manufacturing processes, or failure to comply with regulatory requirements, may result in revisions to the approved labeling to add new safety information; imposition of post-market studies or clinical trials to assess new safety risks; or imposition of distribution or other restrictions under a REMS program. Other potential consequences include, among other things:
•
restrictions on the marketing or manufacturing of the product, complete withdrawal of the product from the market, or voluntary product recalls;
•
fines, warning, or untitled letters or holds on post-approval clinical trials;
•
refusal of the FDA to approve pending applications or supplements to approved applications, or suspension or revocation of product approvals;
•
product seizure or detention, or refusal to permit the import or export of products; or
•
injunctions or the imposition of civil or criminal penalties.
The FDA strictly regulates marketing, labeling, advertising, and promotion of products that are placed on the market. Drugs may be promoted only for the approved indications and in accordance with the provisions of the approved label. However, companies may share truthful and not misleading information that is otherwise consistent with a product’s FDA approved labeling. The FDA and other agencies actively enforce the laws and regulations prohibiting the promotion of off-label uses, and a company that is found to have improperly promoted off-label uses may be subject to significant liability.
In addition, the distribution of prescription pharmaceutical products is subject to the Prescription Drug Marketing Act (“PDMA”), which regulates the distribution of drugs and drug samples at the federal level, and sets minimum standards for the registration and regulation of drug distributors by the states. Both the PDMA and state laws limit the distribution of prescription pharmaceutical product samples and impose requirements to ensure accountability in distribution.
Hatch-Waxman Amendments
Section 505 of the FDCA describes three types of marketing applications that may be submitted to the FDA to request marketing authorization for a new drug. A Section 505(b)(1) NDA is an application that contains full reports of investigations of safety and efficacy. A 505(b)(2) NDA is an application that contains full reports of investigations of safety and efficacy but where at least some of the information required for approval comes from investigations that were not conducted by or for the applicant and for which the applicant has not obtained a right of reference or use from the person by or for whom the investigations were conducted. This regulatory pathway enables the applicant to rely, in part, on the FDA’s prior findings of safety and efficacy for an existing product, or published literature, in support of its application. The FDA may also require companies to perform additional studies or measurements, including clinical trials, to support the change from the approved branded reference drug. The FDA may then approve the new product candidate for all, or some, of the labeled indications for which the branded reference drug has been approved, as well as for any new indication sought by the 505(b)(2) applicant. Section 505(j) establishes an abbreviated approval process for a generic version of approved drug products through the submission of an Abbreviated New Drug Application (“ANDA”). An ANDA provides for marketing of a generic drug product that has the same active ingredients, dosage form, strength, route of administration, labeling, performance characteristics, and intended use, among other things, to a previously approved product, known as a reference listed drug (“RLD”). ANDAs are termed “abbreviated” because they are generally not required to include preclinical (animal) and clinical (human) data to establish safety and efficacy. Instead, generic applicants must scientifically demonstrate that their product is bioequivalent to, or performs in the same manner as, the innovator drug through in vitro, in vivo, or other testing. In certain situations, an applicant may obtain ANDA
approval of a generic product with a strength or dosage form that differs from a referenced innovator drug pursuant to the filing and approval of an ANDA Suitability Petition. The FDA will approve the generic product as suitable for an ANDA application if it finds that the generic product does not raise new questions of safety and effectiveness as compared to the innovator product.
Non-Patent Exclusivity
Under the Hatch-Waxman Amendments, the FDA may not approve (or in some cases accept) an ANDA or 505(b)(2) application until any applicable period of non-patent exclusivity for the RLD has expired. The FDCA provides a period of five years of non-patent data exclusivity for a new drug containing a new chemical entity (“NCE”). For the purposes of this provision, an NCE is a drug that contains no active moiety that has previously been approved by the FDA in any other NDA. An active moiety is the molecule or ion responsible for the physiological or pharmacological action of the drug substance. In cases where such NCE exclusivity has been granted, an ANDA may not be filed with the FDA until the expiration of five years unless the submission is accompanied by a Paragraph IV certification, which states the proposed generic drug will not infringe one or more of the already approved product’s listed patents or that such patents are invalid or unenforceable, in which case the applicant may submit its application four years following the original product approval.
The FDCA also provides for a period of three years of exclusivity for non-NCE drugs if the NDA or a supplement to the NDA includes reports of one or more new clinical investigations, other than bioavailability or bioequivalence studies, that were conducted by or for the applicant and are essential to the approval of the application or supplement. This three-year exclusivity period often protects changes to a previously approved drug product, such as a new dosage form, route of administration, combination, or indication, but it generally would not protect the original, unmodified product from generic competition. Unlike five-year NCE exclusivity, an award of three-year exclusivity does not block the FDA from accepting ANDAs seeking approval for generic versions of the drug as of the date of approval of the original drug product; it only prevents FDA from approving such ANDAs.
A drug product can obtain pediatric market exclusivity in the United States. Pediatric exclusivity, if granted, adds six months to existing exclusivity periods for all formulations, dosage forms, and indications of the active moiety and to patent terms. This six-month exclusivity, which runs from the end of other exclusivity protection and patent term, may be granted based on the voluntary completion of a pediatric study in accordance with an FDA-issued “Written Request” for such a study, provided that at the time pediatric exclusivity is granted there is not less than nine months of term remaining.
Hatch-Waxman Patent Certification and the 30-Month Stay
In seeking approval of an NDA or a supplement thereto, NDA sponsors are required to list with the FDA each patent with claims that cover the applicant’s product or an approved method of using the product. Upon approval of an NDA, each of the patents listed in the application for the drug is published in the FDA’s Approved Drug Products with Therapeutic Equivalence Evaluations, commonly known as the Orange Book. Upon submission of an ANDA or 505(b)(2) NDA, an applicant is required to certify to the FDA concerning any patents listed for the RLD in the Orange Book that:
•
no patent information on the drug product that is the subject of the application has been submitted to the FDA;
•
such patent has expired;
•
the date on which such patent expires; or
•
such patent is invalid, unenforceable, or will not be infringed upon by the manufacture, use, or sale of the drug product for which the application is submitted.
Generally, the ANDA or 505(b)(2) NDA cannot be approved until all listed patents have expired, except where the ANDA or 505(b)(2) NDA applicant challenges a listed patent through the last type of certification, also known as a paragraph IV certification. If the applicant does not challenge the listed patents or indicates that it is not seeking approval of a patented method of use, the ANDA or 505(b)(2) NDA application will not be approved until all of the listed patents claiming the referenced product have
expired. If the ANDA or 505(b)(2) NDA applicant has provided a paragraph IV certification, the applicant must send notice of the paragraph IV certification to the NDA and patent holders once the application has been accepted for filing by the FDA. The NDA and patent holders may then initiate a patent infringement lawsuit in response to the notice of the paragraph IV certification. If the paragraph IV certification is challenged by an NDA holder or the patent owner(s) asserts a patent challenge to the paragraph IV certification, the FDA may not approve that application until the earlier of 30 months from the receipt of the notice of the paragraph IV certification, the expiration of the patent, when the infringement case concerning each such patent was favorably decided in the applicant’s favor or settled, or such shorter or longer period as may be ordered by a court. This prohibition is generally referred to as the 30-month stay. In instances where an ANDA or 505(b)(2) NDA applicant files a paragraph IV certification, the NDA holder or patent owner(s) regularly take action to trigger the 30-month stay, recognizing that the related patent litigation may take many months or years to resolve. Thus, approval of an ANDA or 505(b)(2) NDA could be delayed for a significant period of time depending on the patent certification the applicant makes and the reference drug sponsor’s decision to initiate patent litigation. If the drug has NCE exclusivity and the ANDA is submitted four years after approval, the 30-month stay is extended so that it expires seven and a half years after approval of the innovator drug, unless the patent expires or there is a decision in the infringement case that is favorable to the ANDA applicant before then.
Patent Term Restoration and Extension
A patent claiming a new drug product may be eligible for a limited patent term extension under the Hatch-Waxman Amendments, which permits a patent term restoration of up to five years for patent term lost during product development and the FDA regulatory review. The restoration period granted is typically one-half the time between the effective date of an IND and the submission date of an NDA, plus the time between the submission date of an NDA and the ultimate approval date, provided the sponsor acted with diligence. Patent term restoration cannot be used to extend the remaining term of a patent past a total of 14 years from the product’s approval date. Only one patent applicable to an approved drug product is eligible for the extension, and the application for the extension must be submitted prior to the expiration of the patent in question and within 60 days of drug approval. A patent that covers multiple drugs for which approval is sought can only be extended in connection with one of the approvals. The U.S. Patent and Trademark Office (“USPTO”) reviews and approves the application for any patent term extension or restoration in consultation with the FDA.
Rest of the World Regulation
For other countries outside of the United States, including those in Europe, Latin America, or Asia, the requirements governing product development, the conduct of clinical trials, product marketing, product licensing, pricing and reimbursement can vary from country to country. Failure to comply with applicable foreign regulatory requirements may subject sponsors, manufacturers, or marketers of pharmaceutical products to, among other things, fines, suspension, or withdrawal of regulatory authorizations and approvals, product recalls, seizure of products, operating restrictions and criminal prosecution.
Review and Approval of Medicinal Products in the European Union
In order to market any product outside of the United States, a company must also comply with numerous and varying regulatory requirements of other countries and jurisdictions regarding quality, safety, and efficacy, and governing, among other things, clinical trials, obtaining marketing authorization (“MA”), commercial sales and distribution of products. Whether or not it obtains FDA approval for a product, an applicant will need to obtain the necessary approvals by the comparable foreign regulatory authorities before it can commence clinical trials or marketing of the product in those countries or jurisdictions. Specifically, the process governing approval of medicinal products in the EU generally follows similar lines as in the United States. It entails satisfactory completion of preclinical studies and adequate and well-controlled clinical trials to establish the safety and efficacy of the product for each proposed indication. It also requires the submission to the relevant competent authorities of an MA application (“MAA”), and granting of an MA by these authorities before the product can be marketed and sold in the EU.
Non-clinical Studies and Clinical Trials
Similarly to the United States, the various phases of non-clinical and clinical research in the EU are subject to significant regulatory controls.
Non-clinical studies are performed to demonstrate the health or environmental safety of new chemical or biological substances. Non-clinical (pharmaco-toxicological) studies must be conducted in compliance with GLP, as set forth in EU Directive 2004/10/EC (unless otherwise justified for certain particular medicinal products, e.g., radio-pharmaceutical precursors for radio-labeling purposes). These GLP standards reflect the Organization for Economic Co-operation and Development requirements.
Clinical trials of medicinal products in the EU must be conducted in accordance with EU and national regulations and the International Council for Harmonization of Technical Requirements for Pharmaceuticals for Human Use (“ICH”) guidelines on GCP, as well as the applicable regulatory requirements and the ethical principles that have their origin in the Declaration of Helsinki. If the sponsor of the clinical trial is not established within the EU, it must appoint an EU entity to act as its legal representative. The sponsor must take out a clinical trial insurance policy, and in most EU member states, the sponsor is liable to provide ‘no fault’ compensation to any study subject injured in the clinical trial.
The EU Clinical Trials Regulation (“CTR”), which was adopted in April 2014 and repealed the EU Clinical Trials Directive, became applicable on January 31, 2022. Unlike directives, the CTR is directly applicable in all EU member states without the need for member states to further implement it into national law. The CTR notably harmonizes the assessment and supervision processes for clinical trials throughout the EU via a Clinical Trials Information System, which contains a centralized EU portal and database.
While the EU Clinical Trials Directive required a separate clinical trial authorization application (“CTA”) to be submitted in each member state in which the clinical trial takes place, to both the competent national health authority and an independent ethics committee, much like the FDA and IRB respectively, the CTR introduces a centralized process and only requires the submission of a single application for multi-center trials. The CTR allows sponsors to make a single submission to both the competent authority and an ethics committee in each member state, leading to a single decision per member state. The CTA must include, among other things, a copy of the trial protocol and an investigational medicinal product dossier containing information about the manufacture and quality of the medicinal product under investigation. The assessment procedure of the CTA has been harmonized as well, including a joint assessment by all member states concerned, and a separate assessment by each member state with respect to specific requirements related to its own territory, including ethics rules. Each member state’s decision is communicated to the sponsor via the centralized Clinical Trials Information System. Once the CTA is approved, clinical study development may proceed.
Medicines used in clinical trials must be manufactured in accordance with Good Manufacturing Practice (“GMP”). Other national and EU-wide regulatory requirements may also apply.
Marketing Authorization
In order to market our product candidates in the EU and many other foreign jurisdictions, we must obtain separate regulatory approvals. More concretely, in the EU, medicinal product candidates can only be commercialized after obtaining an MA. To obtain an MA, an applicant must submit an MAA either under a centralized procedure administered by the EMA, or one of the procedures administered by competent authorities in the EU member states (decentralized procedure or mutual recognition procedure) for obtaining an MA in multiple EU member states.
“Centralized MAs” are issued by the European Commission through the centralized procedure based on the opinion of the Committee for Medicinal Products for Human Use (“CHMP”) of the EMA, and are valid throughout the EU. Pursuant to Regulation (EC) No 726/2004, the centralized procedure is compulsory for specific products, including for therapeutics produced by certain biotechnological processes, products designated as orphan medicinal products, advanced therapy medicinal products
(gene therapy, somatic cell therapy, and tissue-engineered products) and products with a new active substance indicated for the treatment of certain diseases, including products for the treatment of HIV, AIDS, cancer, diabetes, neurodegenerative diseases, auto-immune and other immune dysfunctions and viral diseases. The centralized procedure is optional for products containing a new active substance not yet authorized in the EU, or for products that constitute a significant therapeutic, scientific, or technical innovation or which are in the interest of public health in the EU.
Under the centralized procedure, the maximum timeframe for the evaluation of an MAA by the EMA is 210 days, excluding clock stops, when additional information or written or oral explanation is to be provided by the applicant in response to questions asked by the CHMP. Clock stops may extend the timeframe of evaluation of an MAA considerably beyond 210 days. Accelerated evaluation might be granted by the CHMP in exceptional cases, when a medicinal product is of major interest from a public health perspective and in particular from the point of view of therapeutic innovation. If the CHMP accepts such request, the time limit of 210 days will be reduced to 150 days, excluding clock stops, but it is possible that the CHMP can revert to the standard time limit for the centralized procedure if it considers that it is no longer appropriate to conduct an accelerated assessment. At the end of this period, the CHMP provides a scientific opinion on whether or not a marketing authorization should be granted in relation to a medicinal product. Within 67 days from the date of the CHMP opinion, the European Commission will adopt its final decision on the MAA.
“National MAs” are issued by the competent authorities of the EU member states, only cover their respective territory, and are available for product candidates not falling within the mandatory scope of the centralized procedure. Where a product has already been authorized for marketing in an EU member state, this national MA can be recognized in another member state through the mutual recognition procedure. If the product has not received a national MA in any member state at the time of application, it can be approved simultaneously in various member states through the decentralized procedure. Under the decentralized procedure an identical dossier is submitted to the competent authorities of each of the member states in which the MA is sought, one of which is selected by the applicant as the reference member state.
Periods of Authorization and Renewals
An MA has an initial validity of five years. The MA may be renewed after five years on the basis of a re-evaluation of the risk-benefit balance by the EMA or by the competent authority of the relevant EU member state for a nationally authorized product. To this end, the MA holder must provide the EMA or the competent authority with a consolidated version of the file in respect of quality, safety, and efficacy, including all variations introduced since the MA was granted, at least nine months before the MA ceases to be valid. Once renewed, the MA is valid for an unlimited period, unless the European Commission or the competent authorities of the relevant member states decide, on justified grounds relating to pharmacovigilance, to proceed with one further five year renewal period. Any authorization which is not followed by the actual placing of the medicinal product on the EU market (for centrally-authorized products) or on the market of the authorizing EU member state (for nationally-authorized products) within three years after authorization ceases to be valid (the so-called “sunset clause”).
Data and Market Exclusivity
In the EU, innovative medicinal products approved on the basis of a complete and independent data package generally receive eight years of data exclusivity and an additional two years of market exclusivity upon grant of an MA. The data exclusivity period prevents applicants for authorization of generics or biosimilars of these innovative products from referencing the innovator’s preclinical and clinical trial data contained in the dossier of the reference product when applying for a generic or biosimilar (abbreviated) MA, for a period of eight years from the date on which the reference product was first authorized in the EU. During an additional two-year period of market exclusivity, a generic or biosimilar MAA can be submitted, and the innovator’s data may be referenced, but no generic or biosimilar medicinal product can be placed on the EU market until the expiration of the market exclusivity. The overall 10-year period will be extended to a maximum of 11 years if, during the first eight years of those 10 years, the MA holder obtains an authorization for one or more new therapeutic indications which,
during the scientific evaluation prior to their authorization, are held to bring a significant clinical benefit in comparison with existing therapies. There is no guarantee that a product will be considered by the EMA to be an innovative medicinal product, and products may not qualify for data exclusivity. Even if a product is considered to be an innovative medicinal product so that the innovator gains the prescribed period of data exclusivity, another company nevertheless could also market another version of the product if such company obtained an MA based on an MAA with a complete and independent data package of pharmaceutical tests, preclinical tests and clinical trials.
Orphan Medicinal Products
The criteria for designating an “orphan medicinal product” in the EU are similar in principle to those in the United States. A medicinal product can be designated as an orphan if its sponsor can establish that: (1) the product is intended for the diagnosis, prevention, or treatment of a life-threatening or chronically debilitating condition, (2) either (i) such condition affects no more than five in ten thousand persons in the EU when the application is made, or (ii) without the benefits derived from orphan status, it is unlikely that the marketing of the product in the EU would generate sufficient return to justify the necessary investment in its development; (3) there exists no satisfactory method of diagnosis, prevention, or treatment of the condition in question that has been authorized in the EU or, if such method exists, the product would be of significant benefit to those affected by that condition.
An orphan designation provides a number of benefits, including fee reductions, regulatory assistance, and the possibility to apply for a centralized EU MA. Upon grant of an MA, orphan medicinal products are entitled to a ten-year period of market exclusivity, which means that the EMA and the competent authorities of the EU member states cannot accept another MAA, or grant an MA, or accept an application to extend an MA for a similar medicinal product for the same indication for a period of ten years. A “similar medicinal product” is defined as a medicinal product containing a similar active substance or substances as contained in an authorized orphan medicinal product, and which is intended for the same therapeutic indication. The period of market exclusivity may be extended by two years for orphan medicinal products that have also complied with an agreed pediatric investigation plan (“PIP”). No extension to any supplementary protection certificate (“SPC”) can be granted on the basis of pediatric studies for orphan indications. Orphan designation does not convey any advantage in, or shorten the duration of, the regulatory review and approval process.
The orphan exclusivity period may be reduced to six years if, at the end of the fifth year, it is established that the product no longer meets the criteria for which it received orphan designation, including where it is shown that the product is sufficiently profitable not to justify maintenance of market exclusivity or where the prevalence of the condition has increased above the threshold. Additionally, an MA may be granted to a similar product for the same indication as an authorized orphan product at any time if (i) a second applicant can establish that its product, although similar to the authorized orphan product, is safer, more effective or otherwise clinically superior; (ii) the MA holder for the authorized orphan product consents to a second medicinal product application; or (iii) the MA holder for the authorized product cannot supply enough orphan medicinal product.
Pediatric Development
Regulation (EC) No 1901/2006 provides that prior to obtaining an MA in the EU, applicants have to demonstrate compliance with all measures included in a PIP, agreed with the EMA’s Pediatric Committee (“PDCO”), and covering all subsets of the pediatric population, unless the PDCO has granted (1) a product-specific waiver, (2) a class waiver, or (3) a deferral for one or more of the measures included in the PIP. The PIP sets out the timing and measures proposed to generate data to support a pediatric indication of the product for which an MA is being sought. Products that are granted an MA with the results of the pediatric clinical trials conducted in accordance with the PIP are eligible for a six-month extension of the protection under an SPC provided an application for such extension is made at the same time as filing the SPC application for the product, or at any point up to two years before the SPC expires, even where the trial results are negative. In the case of orphan medicinal products, a two-year extension of the orphan market exclusivity may be available. This pediatric reward is subject to specific conditions and is not automatically available when data in compliance with the PIP are developed and submitted.
Post-Approval Requirements
Similar to the United States, both MA holders and manufacturers of medicinal products are subject to comprehensive regulatory oversight by the EMA, the European Commission and/or the competent regulatory authorities of the member states. The holder of an MA must establish and maintain a pharmacovigilance system and appoint an individual qualified person for pharmacovigilance (“QPPV”), who is responsible for the establishment and maintenance of that system, and oversees the safety profiles of medicinal products and any emerging safety concerns. Key obligations include expedited reporting of suspected serious adverse reactions and submission of periodic safety update reports (“PSURs”).
All new MAAs must include a risk management plan (“RMP”), describing the risk management system that the company will put in place and documenting measures to prevent or minimize the risks associated with the product.
Regulatory Authorities may also impose specific obligations as a condition of the MA. Such risk-minimization measures or post-authorization obligations may include additional safety monitoring, more frequent submission of PSURs, or the conduct of additional clinical trials or post-authorization safety studies.
The advertising and promotion of medicinal products is also subject to laws concerning promotion of medicinal products, interactions with physicians, misleading and comparative advertising, and unfair commercial practices. All advertising and promotional activities for the product must be consistent with the approved summary of product characteristics, and therefore all off-label promotion is prohibited in the EU. Direct-to-consumer advertising of prescription medicines is also prohibited in the EU. Although general requirements for advertising and promotion of medicinal products are established under EU directives, the details are governed by regulations in each member state and can differ from one country to another.
The aforementioned EU rules are generally applicable in the European Economic Area (“EEA”), which consists of the EU member states plus Norway, Liechtenstein, and Iceland.
Failure to comply with EU and member state laws that apply to the conduct of clinical trials, manufacturing approval, authorization of medicinal products and marketing of such products, both before and after grant of the MA, manufacturing of pharmaceutical products, statutory health insurance, bribery and anti-corruption, or with other applicable regulatory requirements may result in administrative, civil, or criminal penalties. These penalties could include delays or refusal to authorize the conduct of clinical trials, or to grant an MA, product withdrawals and recalls, product seizures, suspension, withdrawal or variation of the MA, total or partial suspension of production, distribution, manufacturing, or clinical trials, operating restrictions, injunctions, suspension of licenses, fines, and criminal penalties.
Reform of the Regulatory Framework in the European Union
The European Commission introduced legislative proposals in April 2023 intended to replace the current regulatory framework in the EU for all medicines (including those for rare diseases and for children). In April 2024, the European Parliament adopted its position on the legislative proposals and, in June 2025, the Council of the European Union adopted its position. A common position on the text was agreed upon on December 11, 2025, in the context of subsequent inter-institutional trilogue negotiations. The proposed revisions remain to be formally adopted into EU law, and are not expected to become applicable before 2028.
Brexit and the Regulatory Framework in the United Kingdom
Following the end of the Brexit transition period on January 1, 2021 and the implementation of the Windsor Framework on January 1, 2025, the United Kingdom (“UK”) is not generally subject to EU laws in respect of medicines. The EU laws that have been transposed into UK law through secondary legislation remain applicable in the UK; however, new legislation such as the (EU) CTR is not applicable in the UK.
Under the Medicines and Medical Devices Act 2021, the Secretary of State or an ‘appropriate authority’ have delegated powers to amend or supplement existing regulations in the area of medicinal products and medical devices. This allows new rules to be introduced in the future by way of secondary legislation, which aims to allow flexibility in addressing regulatory gaps and future changes in the fields of human medicines, clinical trials, and medical devices.
As of January 1, 2021, the Medicines and Healthcare products Regulatory Agency (“MHRA”) is the UK’s standalone medicines and medical devices regulator. As a result of the Northern Ireland Protocol, different rules initially applied in Northern Ireland than in England, Wales, and Scotland (together, “Great Britain”, or GB), as Northern Ireland continued to follow the EU regulatory regime for a period of time following Brexit. However, on January 1, 2025 a new arrangement called the “Windsor Framework” came into effect and reintegrated Northern Ireland under the regulatory authority of the MHRA with respect to medicinal products. The Windsor Framework removes EU licensing processes and EU labeling and serialization requirements in relation to Northern Ireland and introduces a UK-wide licensing process for medicines.
The UK regulatory framework in relation to clinical trials is governed by the Medicines for Human Use (Clinical Trials) Regulations 2004, as amended, which is derived from the Clinical Trials Directive, as implemented into UK national law through secondary legislation. In April 2025, the UK introduced the Medicines for Human Use (Clinical Trials) (Amendment) Regulations, with the aim of creating a streamlined, risk-proportionate system that accelerates approvals while maintaining robust safety standards. Following a 12-month implementation period, the amended regulations came fully into force on 28 April 2026.
In addition, in October 2023, the MHRA announced a new Notification Scheme for clinical trials which enables a more streamlined and risk-proportionate approach to initial clinical trial applications for Phase 4 and low-risk Phase 3 clinical trial applications.
MAs in the UK are governed by the Human Medicines Regulations (SI 2012/1916), as amended. All existing EU MAs for centrally authorized products were automatically converted or grandfathered into UK MAs, effective in GB (only), free of charge on January 1, 2021, unless the MA holder chose to opt-out. Under the terms of the Windsor Framework, these licenses became valid for the whole of the UK from January 1, 2025. In order to use the centralized procedure to obtain an MA that will be valid throughout the EEA, companies must be established in the EEA. Therefore, after Brexit, companies established in the UK can no longer use the EU centralized procedure and instead an EEA entity must hold any centralized MAs. In order to obtain a UK MA to commercialize products in the UK, an applicant must follow one of the UK national authorization procedures or one of the remaining post-Brexit international cooperation procedures. The MHRA has introduced changes to national licensing procedures, including procedures to prioritize access to new medicines that will benefit patients, a 150-day assessment (subject to clock-stops) and a rolling review procedure. The rolling-review procedure permits the separate or joint submission of quality, non-clinical, and clinical data to the MHRA which can be reviewed on a rolling basis. After an application under the rolling-review procedure has been validated, the final decision should be received within 100 days (subject to clock-stops). In addition, since January 1, 2024, the MHRA may rely on the International Recognition Procedure (“IRP”), when reviewing certain types of MAAs. Pursuant to the IRP, the MHRA will take into account the expertise and decision-making of trusted regulatory partners (e.g. the medicines regulatory authorities in Australia, Canada, Switzerland, Singapore, Japan, the U.S.A. and the EMA in the EU) when considering an application for a UK MA. The MHRA will conduct a targeted assessment of IRP applications but retain the authority to reject applications if the evidence provided is considered insufficiently robust. Applications should be decided within a maximum of 60 days if there are no major objections identified that cannot be resolved within such 60 day period and the approval from the trusted regulatory partner selected has been granted within the previous 2 years. If there are such major objections identified or such approval hasn’t been granted within the previous 2 years, then the relevant timeframe for the MHRA decision is within 110 days. Applicants can submit initial MAAs to the IRP but the procedure can also be used throughout the lifecycle of a product for post-authorization procedures including line extensions, variations, and renewals.
In the UK, the initial duration of an MA is five years and following renewal will be valid for an unlimited period unless the MHRA decides on justified grounds relating to pharmacovigilance, to proceed with only one additional five-year renewal. Any authorization which is not followed by the actual placing of the medicine on the market in the UK (or GB, if the MA is only valid in GB) within three (3) years shall cease to be in force.
There is no pre-MA orphan designation in the UK. Instead, the MHRA reviews applications for orphan designation in parallel to the corresponding MA application. The criteria are essentially the same, but have been tailored for the market, i.e., the prevalence of the condition in the UK, rather than the EU, must not be more than five in 10,000. Should an orphan designation be granted, the period of market exclusivity will be set from the date of first approval of the product in the UK.
Review and Approval of Medicinal Products in Australia
The Therapeutic Goods Administration (“TGA”) and the National Health and Medical Research Council (“NHMRC”) set the GCP requirements for clinical research in Australia.
Compliance with the regulations, standards and codes set by the TGA and NHMRC is mandatory. Under the Therapeutic Goods Act 1989 (Cth) and the Therapeutic Goods Regulations 1990 (Cth), it is a condition (amongst other conditions) of all clinical trials involving investigational medicinal products to comply with the National Statement on Ethical Conduct in Research Involving Humans, published by the NHMRC (the National Statement), and the Guideline for Good Clinical Practice published by the International Council for Harmonization of Technical Requirements for Registration of Pharmaceuticals for Human Use (“ICH Guidelines”). The ICH Guidelines have been adopted in Australia, and must be complied with across all fields of clinical research involving therapeutic goods, including those related to pharmaceutical quality, nonclinical and clinical data requirements and trial designs. The basic requirements for preclinical data to support a first-in-human trial under ICH Guidelines are applicable in Australia. Requirements related to adverse event reporting in Australia are generally similar to those required in other major jurisdictions (and there is alignment with the European Union’s Clinical Trial Regulations: Regulation EU No 536/2014), although reporting timeframes may differ to other jurisdictions.
Clinical trials conducted using “unapproved therapeutic goods” in Australia, being those which have not yet been evaluated by the TGA for quality, safety and efficacy (and including unapproved indications of therapeutic goods which have otherwise been approved for use in Australia) must occur pursuant to either the Clinical Trial Notification Scheme (“CTN Scheme”) or the Clinical Trial Approval Scheme (“CTA Scheme”). In each case, the trial is supervised by a Human Research Ethics Committee (“HREC”), an independent review committee constituted in accordance with the National Statement that ensures the protection of rights, safety and well-being of human subjects involved in a clinical trial. An HREC reviews, approves and provides continuing oversight of trial protocols (including any amendments), methods and materials intended to be used in obtaining and documenting informed consent of the clinical trial subjects.
The CTN Scheme broadly involves:
•
the investigator or sponsor of the Australian clinical trial submitting a ‘Notification of Intent to Conduct a Clinical Trial’ form (“CTN Form”) to the TGA and payment of the relevant fee;
•
the TGA may request further specific information relating to the ‘unapproved therapeutic goods’ that are the subject of the clinical trial;
•
submission to an HREC, of all material relating to the proposed clinical trial, including the trial protocol;
•
the HREC reviews the scientific validity of the trial design, the balance of risk versus harm of the therapeutic good, the ethical acceptability of the trial process, and approves the trial protocol. The HREC is also responsible for monitoring the conduct of the trial;
•
the institution or organization at which the trial will be conducted, referred to as the “Approving Authority,” giving final approval for the conduct of the trial at the site, in terms no less restrictive to those advised by the HREC; and
•
ensuring that the CTN form is signed by the sponsor, the principal investigator, the chairman of the HREC and a person responsible from the Approving Authority. The TGA does not review any data relating to the clinical trial, however CTN trials cannot commence until the trial has been notified to the TGA. It is the responsibility of the sponsor to ensure that all relevant approvals are in place before supplying the ‘unapproved’ therapeutic goods in the clinical trial in Australia.
Under the CTA Scheme:
•
a sponsor submits an application to conduct a clinical trial to the TGA for evaluation and comment, which includes payment of the relevant fees. The TGA encourages all sponsors to request a pre-submission meeting with the TGA in order to clarify any questions about existing studies or the proposed data package for the CTA application, and obtain specific advice
•
from the TGA relating to the CTA application process, including the best ways to submit the application and dossier;
•
the TGA will undertake a preliminary assessment to ensure that there is sufficient data to begin evaluation. If critical data is missing, the TGA may request further information;
•
a sponsor must forward any comments made by the TGA Delegate to the HREC(s) at the sites where the trial will be conducted;
•
the HREC is responsible for considering the scientific and ethical issues of the proposed trial protocol.
A sponsor cannot commence a trial under the CTA Scheme until written advice has been received from the TGA regarding the application and approval for the conduct of the trial has been obtained from an ethics committee and the institution at which the trial will be conducted.
Approval for inclusion in the Australian Register of Therapeutic Goods (“ARTG”), is required before a therapeutic good (including pharmaceutical product) may be marketed (or supplied, imported, exported or manufactured) in Australia. Exceptions apply to therapeutic goods/pharmaceutical products that are supplied, imported, and exported to and from Australia for the purposes of a clinical trial, on the basis that certain conditions are met (e.g., the trial is conducted in accordance with the CTN or CTA scheme).
Once a sponsor decides to register a therapeutic good/pharmaceutical product in Australia, in order to obtain registration of the product on the ARTG, it is required that (amongst others):
•
the sponsor submits appropriate documentation, including the outcomes of clinical trials and studies, to allow the TGA to assess the quality, safety and efficacy of the therapeutic product/ pharmaceutical product; and
•
the sponsor submits evidence which demonstrates that the manufacture of the therapeutic product/pharmaceutical product complies with the applicable GMP requirements.
The TGA has the ultimate discretion to decide whether to include the therapeutic product/ pharmaceutical product in the ARTG.
Coverage and Reimbursement
The success of our product candidates, if approved, depends on the availability of coverage and adequate reimbursement from third-party payors. We cannot be sure that coverage and reimbursement will be available for our product candidates, nor can we accurately estimate the potential revenue from them. Similarly, we cannot assure that coverage and reimbursement will be available for any product that we may develop.
In the United States, patients who are provided medical treatment for their conditions generally rely on third-party payors to reimburse all or part of the costs associated with their treatment. Coverage and adequate reimbursement from governmental healthcare programs, such as Medicare and Medicaid, and commercial payors are critical to new product acceptance.
Government authorities and other third-party payors, such as private health insurers and health maintenance organizations, decide which drugs and treatments they will cover and the amount of reimbursement. Coverage and reimbursement by a third-party payor may depend upon a number of factors, including the third-party payor’s determination that the use of a product is:
•
a covered benefit under its health plan;
•
safe, effective and medically necessary;
•
appropriate for the specific patient;
•
cost-effective; and
•
neither experimental nor investigational.
In the United States, no uniform policy of coverage and reimbursement for products exists among third-party payors. As a result, obtaining coverage and reimbursement approval of a product from a government or other third-party payor is a time-consuming and costly process that could require us to provide to each payor supporting scientific, clinical, and cost-effectiveness data for the use of our products on a payor-by-payor basis, with no assurance that coverage and adequate reimbursement will be obtained. In the United States, the principal decisions about reimbursement for new medicines are typically made by the Centers for Medicare and Medicaid Services (“CMS”). CMS decides whether and to what extent a new medicine will be covered and reimbursed under Medicare and private payors tend to follow CMS to a substantial degree. Even if we obtain coverage for a given product, the resulting reimbursement payment rates might not be adequate for us to achieve or sustain profitability or may require co-payments that patients find unacceptably high. Additionally, third-party payors may not cover, or provide adequate reimbursement for, long-term follow-up evaluations required following the use of product candidates, once approved. Patients are unlikely to use our product candidates, once approved, unless coverage is provided and reimbursement is adequate to cover a significant portion of their cost. There is significant uncertainty related to insurance coverage and reimbursement of newly approved products. It is difficult to predict at this time what third-party payors will decide with respect to the coverage and reimbursement for our product candidates.
Net prices for drugs may be reduced by mandatory discounts or rebates required by government healthcare programs or private payors and by any future relaxation of laws that presently restrict imports of drugs from countries where they may be sold at lower prices than in the United States. Increasingly, third-party payors are requiring that drug companies provide them with predetermined discounts from list prices and are challenging the prices charged for medical products. We cannot be sure that reimbursement will be available for any product candidate that we commercialize and, if reimbursement is available, the level of reimbursement. In addition, many pharmaceutical manufacturers must calculate and report certain price reporting metrics to the government, such as average sales price and best price. Penalties may apply in some cases when such metrics are not submitted accurately and timely. Further, these prices for drugs may be reduced by mandatory discounts or rebates required by government healthcare programs. Payment methodologies may be subject to changes in healthcare legislation and regulatory initiatives.
Moreover, increasing efforts by governmental and other third-party payors in the United States and abroad to cap or reduce healthcare costs may cause such organizations to limit both coverage and the level of reimbursement for newly approved products and, as a result, they may not cover or provide adequate payment for our product candidates. There has been increasing legislative and enforcement interest in the United States with respect to specialty drug pricing practices. Specifically, there have been several recent U.S. Congressional inquiries and proposed and enacted federal and state legislation designed to, among other things, bring more transparency to drug pricing, reduce the cost of prescription drugs under Medicare, review the relationship between pricing and manufacturer patient programs and reform government program reimbursement methodologies for drugs.
At the state level, legislatures have increasingly passed legislation and implemented regulations designed to control pharmaceutical and biological product pricing, including price or patient reimbursement constraints, discounts, restrictions on certain product access and marketing cost disclosure and
transparency measures, and, in some cases, designed to encourage importation from other countries and bulk purchasing.
We expect that healthcare reform measures that may be adopted in the future may result in more rigorous coverage criteria and in additional downward pressure on the price that we receive for any approved product. The implementation of cost containment measures or other healthcare reforms may prevent us from being able to generate revenue, attain profitability, or commercialize our products. Legislative and regulatory proposals have been made to expand post-approval requirements and restrict sales and promotional activities for pharmaceutical products. We cannot be sure whether additional legislative changes will be enacted, or whether existing regulations, guidance or interpretations will be changed, or what the impact of such changes on the marketing approvals or clearances of our product candidates, if any, may be.
In addition, in some foreign countries, the proposed pricing for a drug must be approved before it may be lawfully marketed. The requirements governing drug pricing vary widely from country to country. For example, the EU provides options for its Member States to restrict the range of medicinal products for which their national health insurance systems provide reimbursement and to control the prices of medicinal products for human use. To obtain reimbursement or pricing approval, some of these countries may require the completion of clinical trials that compare the cost effectiveness of a particular product candidate to currently available therapies. A Member State may approve a specific price for the medicinal product or it may instead adopt a system of direct or indirect controls on the profitability of the company placing the medicinal product on the market. There can be no assurance that any country that has price controls or reimbursement limitations for pharmaceutical products will allow favorable reimbursement and pricing arrangements for any of our product candidates. Historically, products launched in the EU do not follow price structures of the United States and generally prices tend to be significantly lower.
Healthcare Reform and Other Regulatory Changes
In the United States and some foreign jurisdictions, there have been, and likely will continue to be, a number of legislative and regulatory changes and proposed changes regarding the healthcare system directed at broadening the availability of healthcare, improving the quality of healthcare, and containing or lowering the cost of healthcare.
The Patient Protection and Affordable Care Act, as amended by the Health Care and Education Reconciliation Act (each as amended, collectively known as the “ACA”), which was enacted in 2010, substantially changed the way healthcare is financed by both governmental and private insurers in the United States, and significantly affected the pharmaceutical industry. The ACA contains a number of provisions of particular import to the pharmaceutical and biotechnology industries, including, but not limited to, those governing enrollment in federal healthcare programs. Since its enactment, there have been judicial and Congressional challenges to certain aspects of the ACA, and we expect there will be additional challenges and amendments to the ACA in the future.
Other legislative changes that have been proposed and adopted in the United States since the ACA was enacted include the following:
•
The Budget Control Act of 2011, among other things, created measures for spending reductions by Congress. This includes aggregate reductions of Medicare payments to providers of 2% per fiscal year. Subsequent legislation extended the 2% reduction which remains in effect through 2032.
•
On March 11, 2021, President Biden signed the American Rescue Plan Act of 2021 into law, which eliminated the statutory Medicaid drug rebate cap, previously set at 100% of a drug’s average manufacturer price, for single source and innovator multiple source drugs, effective January 1, 2024. These laws and regulations may result in additional reductions in Medicare and other healthcare funding and otherwise affect the prices we may obtain for any of our product candidates for which we may obtain regulatory approval or the frequency with which any such product candidate is prescribed or used.
Additionally, the Inflation Reduction Act of 2022 (the “IRA”) included several provisions that may impact our business to varying degrees, including provisions that reduce the out-of-pocket spending cap for Medicare Part D beneficiaries from $7,050 to $2,000 starting in 2025, thereby effectively eliminating the coverage gap; impose new manufacturer financial liability on certain drugs under Medicare Part D, allow the U.S. government to negotiate Medicare Part B and Part D price caps beginning January 1, 2026 for certain high-cost drugs and biologics without generic or biosimilar competition; require companies to pay rebates to Medicare for certain drug prices that increase faster than inflation; and delay until January 1, 2032 the implementation of the Department of Health and Human Services’ rebate rule that would have limited the fees that pharmacy benefit managers can charge. Further, under the IRA, orphan drugs are exempted from the Medicare drug price negotiation program, but only if they have one orphan designation and for which the only approved indication is for that disease or condition. Under the One Big Beautiful Bill Act of 2025, this restriction was eliminated; and effective for the 2028 initial price applicability year, all orphan drugs, regardless of the number of orphan drug designations or indications, are exempt from the Medicare drug price negotiation program. Although the effects of the IRA on our business and the healthcare industry in general are not yet fully understood, we are taking into consideration the potential impact of the IRA on our development and commercialization activities.
The cost of prescription drugs and biological products has also been the subject of considerable discussion in the United States. At a federal level, President Trump reversed some of President Biden’s executive orders including rescinding Executive Order 14087 entitled “Lowering Prescription Drug Costs for Americans.” President Trump may issue new executive orders designed to impact prescription product pricing. A number of these and other proposed measures may require authorization through additional legislation to become effective. Congress and the Trump administration have indicated that they will continue to seek new legislative measures to control prescription product costs.
On April 15, 2025, the Trump Administration published Executive Order 14273, “Lowering Drug Prices by Once Again Putting Americans First,” which generally directs the federal government to take measures to reduce drug prices, including eliminating the so-called “pill penalty” under the Inflation Reduction Act that creates a distinction between small molecule and large molecule products for purposes of determining when a drug may be eligible for drug price negotiation. On May 12, 2025, the Trump Administration published Executive Order 14297, “Delivering Most-Favored-Nation Prescription Drug Pricing to American Patients” which generally, among other things, directs the federal government to establish and communicate most-favored-nation (“MFN”) price targets to pharmaceutical manufacturers to bring prices for American patients in line with comparably developed nations. Further, the Executive Order directs the federal government to support regulatory paths to allow direct-to-patient sales for companies that meet these targets. It also states that the Administration will take additional aggressive action (for example, examining whether marketing approvals should be modified or rescinded or opening the door for individual drug importation waivers) should manufacturers fail to offer American consumers the most-favored-nation lowest price. It also directs the Secretary of Commerce and the U.S. Trade Representative to “take all necessary and appropriate action to ensure foreign countries are not engaged in any act, policy, or practice that may be unreasonable or discriminatory or that may impair United States national security . . . including by suppressing the price of pharmaceutical products below fair market value in foreign countries.” Notably, a similar “Most Favored Nation” pricing rule enacted under the first Trump Administration was subject to an injunction resulting from judicial challenges to the rule, which was formally rescinded by the former Biden Administration in August 2021.
On December 19, 2025, CMS released two proposed rules that would incorporate MFN pricing principles into federal reimbursement for prescription drugs: the GLOBE and GUARD models. On October 2, 2026, CMS finalized the GLOBE regulation, effective November 30, 2026, as a mandatory CMS Innovation Center payment model under Section 1115A of the Social Security Act. As finalized, GLOBE will require manufacturers of specified single source drugs and sole source biologics, within designated therapeutic classes and that have incurred more than $100 million of Medicare Part B spending over a 12-month period to pay incremental rebates based on an alternative, internationally benchmarked calculation of the existing Medicare Part B drug inflation rebate. The GLOBE Model will begin on January 1, 2027 with the collection of voluntary manufacturer-submitted international drug net pricing data, and will be tested over 5 performance years, running from April 1, 2027 through
March 31, 2032. The associated payment period will run from April 1, 2027 through March 31, 2034. GLOBE excludes orphan-only drugs, cell and gene therapy products, plasma-derived products, and drugs with an effective Medicare-negotiated maximum fair price.
GUARD remains a proposed regulation that has not been finalized and, as proposed, would similarly mandate manufacturer rebates for qualifying sole source drugs where the Medicare net price exceeds an MFN benchmark derived from international reference pricing methodologies, with its performance period proposed to begin in 2027. The GLOBE model and the proposed GUARD model will likely be subject to legal challenges that could delay implementation or modify their impact on manufacturer pricing and revenue. Additionally, in November 2025, CMS introduced the GENErating cost Reductions fOr U.S. Medicaid (“GENEROUS”) Model, currently proposed as a voluntary MFN framework for manufacturers participating in the Medicaid Drug Rebate Program. Although it is voluntary, the GENEROUS Model could also impact the drug pricing landscape for manufacturers.
In addition, at the state level, legislatures have increasingly passed legislation and implemented regulations similar to those under consideration at the federal level, as well as laws designed to control pharmaceutical and biotherapeutic product pricing, including restrictions on pricing or reimbursement at the state government level, limitations on discounts to patients, marketing cost disclosure and transparency measures, restrictions or other limitations on patient assistance, and, in some cases, policies to encourage importation from other countries (subject to federal approval) and bulk purchasing. Certain states are also pursuing cost containment efforts through Prescription Drug Affordability Boards (“PDABs”) and similar entities. While many PDABs have been granted authority to promote drug price transparency and reporting, some states have granted PDABs more expansive authority, including to set Upper Payment Limits (“UPLs”) on select, high price drugs. The adoption and implementation of UPLs may put downward pressure on drug prices and impact our company’s future revenues.
Other Healthcare Laws and Compliance Requirements
Pharmaceutical companies are subject to additional healthcare regulation and enforcement by the federal government and by authorities in the states and foreign jurisdictions in which they conduct their business that may constrain the financial arrangements and relationships through which we research, as well as sell, market and distribute any products for which we obtain marketing authorization. Arrangements with third-party payors and customers can expose pharmaceutical manufactures to broadly applicable fraud and abuse and other healthcare laws and regulations, including, without limitation, the federal Anti-Kickback Statute and the federal False Claims Act (“FCA”) which may constrain the business or financial arrangements and relationships through which companies research, sell, market and distribute pharmaceutical products. In addition, transparency laws and patient privacy laws can apply to the activities of pharmaceutical manufacturers. The applicable federal, state and foreign healthcare laws and regulations that can affect a pharmaceutical company’s operations include without limitation:
•
The federal Anti-Kickback Statute, which prohibits, among other things, knowingly and willfully soliciting, receiving, offering, or paying any remuneration (including any kickback, bribe, or rebate), directly or indirectly, overtly or covertly, in cash or in kind, to induce, or in return for, either the referral of an individual, or the purchase, lease, order or recommendation of any good, facility, item or service for which payment may be made, in whole or in part, under the Medicare and Medicaid programs, or other federal healthcare programs. A person or entity can be found guilty of violating the statute without actual knowledge of the statute or specific intent to violate it. Violations are subject to civil and criminal fines and penalties for each violation, plus up to three times the remuneration involved, imprisonment, and exclusion from government healthcare programs. In addition, the government may assert that a claim including items or services resulting from a violation of the federal Anti-Kickback Statute constitutes a false or fraudulent claim for purposes of the FCA or federal civil monetary penalties. The Anti-Kickback Statute has been interpreted to apply to arrangements between pharmaceutical manufacturers on the one hand and prescribers, purchasers, and formulary managers on the other. There are a number of statutory exceptions and regulatory safe harbors protecting some
common activities from prosecution, but such exceptions and safe harbors are drawn narrowly and require strict compliance in order to offer protection;
•
The federal civil and criminal false claims laws, including the FCA, and civil monetary penalty laws, which prohibit any person or entity from, among other things, knowingly presenting, or causing to be presented, a false, fictitious, or fraudulent claim for payment to, or approval by, the federal government or knowingly making, using or causing to be made or used a false record or statement, including providing inaccurate billing or coding information to customers or promoting a product off-label, material to a false or fraudulent claim to the federal government. As a result of a modification made by the Fraud Enforcement and Recovery Act of 2009, a claim includes “any request or demand” for money or property presented to the federal government. In addition, manufacturers can be held liable under the FCA even when they do not submit claims directly to government payors if they are deemed to “cause” the submission of false or fraudulent claims. The FCA also permits a private individual acting as a “whistleblower” to bring actions on behalf of the federal government alleging violations of the FCA and to share in any monetary recovery;
•
The federal Health Insurance Portability and Accountability Act of 1996 (“HIPAA”), which created federal criminal statutes that prohibit, among other things, knowingly and willfully executing, or attempting to execute, a scheme to defraud any healthcare benefit program or obtain, by means of false or fraudulent pretenses, representations, or promises, any of the money or property owned by, or under the custody or control of, any healthcare benefit program, regardless of the payor (e.g., public or private) and knowingly and willfully falsifying, concealing or covering up by any trick or device a material fact or making any materially false statements in connection with the delivery of, or payment for, healthcare benefits, items or services relating to healthcare matters. Similar to the federal Anti-Kickback Statute, a person or entity can be found guilty of violating HIPAA without actual knowledge of the statute or specific intent to violate it;
•
HIPAA, as amended by the Health Information Technology for Economic and Clinical Health Act of 2009 (“HITECH”), and their respective implementing regulations, which impose, among other things, specified requirements relating to the privacy, security, and transmission of individually identifiable health information held by covered entities and their business associates as well as their covered subcontractors. HITECH also created new tiers of civil monetary penalties, amended HIPAA to make civil and criminal penalties directly applicable to business associates, and gave state attorneys general new authority to file civil actions for damages or injunctions in federal courts to enforce the federal HIPAA laws and seek attorneys’ fees and costs associated with pursuing federal civil actions;
•
The federal legislation commonly referred to as the Physician Payments Sunshine Act, created under the ACA, and its implementing regulations, which requires certain manufacturers of drugs, devices, biologics and medical supplies for which payment is available under Medicare, Medicaid or the Children’s Health Insurance Program (with certain exceptions) to report annually to CMS, information related to payments or other transfers of value made to physicians, other licensed care professionals, and teaching hospitals, as well as ownership and investment interests held by physicians and their immediate family members;
•
Federal government price reporting laws, which require us to calculate and report complex pricing metrics in an accurate and timely manner to government programs;
•
Federal consumer protection and unfair competition laws, which broadly regulate marketplace activities and activities that potentially harm consumers; and
•
Additionally, we are subject to state and foreign equivalents of each of the healthcare laws and regulations described above, among others, some of which may be broader in scope and may apply regardless of the payor. Many U.S. states have adopted laws similar to the federal Anti-Kickback Statute and False Claims Act, and may apply to our business practices, including, but not limited to, research, distribution, sales, or marketing arrangements and claims involving healthcare items or services reimbursed by non-governmental payors, including
private insurers. In addition, some states have passed laws that require pharmaceutical companies to comply with the April 2003 Office of Inspector General Compliance Program Guidance for Pharmaceutical Manufacturers and/or the Pharmaceutical Research and Manufacturers of America’s Code on Interactions with Healthcare Professionals. Several states also impose other marketing restrictions or require pharmaceutical companies to make marketing or price disclosures to the state and require the registration of pharmaceutical sales representatives. Additionally, some state and local laws require certain regulatory licenses to manufacture or distribute our products commercially and/or the registration of pharmaceutical sales representatives in the jurisdiction. State and foreign laws, including for example the European Union General Data Protection Regulation, which became effective May 2018 also govern the privacy and security of health information in some circumstances, many of which differ from each other in significant ways and often are not preempted by HIPAA, thus complicating compliance efforts. There are ambiguities as to what is required to comply with these state requirements and if we fail to comply with an applicable state law requirement we could be subject to penalties. Finally, there are state and foreign laws governing the privacy and security of health information, many of which differ from each other in significant ways and often are not preempted by HIPAA, thus complicating compliance efforts.
The scope and enforcement of each of these laws is uncertain and subject to rapid change in the current environment of healthcare reform, especially in light of the lack of applicable precedent and regulations with respect to certain laws. Federal and state enforcement bodies have recently increased their scrutiny of interactions between healthcare companies and healthcare providers, which has led to a number of investigations, prosecutions, convictions and settlements in the healthcare industry. Prohibitions or restrictions on sales or withdrawal of future marketed products could materially affect our business in an adverse way. Changes in regulations, statutes or the interpretation of existing regulations could impact our business in the future by requiring, for example: (i) changes to our manufacturing arrangements; (ii) additions or modifications to product labeling; (iii) the recall or discontinuation of our products; or (iv) additional record-keeping requirements. If any such changes were to be imposed, they could adversely affect the operation of our business. Ensuring our business arrangements comply with applicable healthcare laws, as well as responding to possible investigations by government authorities, can be time- and resource-consuming and can divert a company’s attention from the business.
The failure to comply with any of these laws or regulatory requirements subjects companies to possible legal or regulatory action. Depending on the circumstances, failure to meet applicable regulatory requirements can result in significant penalties, including civil, criminal, and administrative penalties, damages, fines, disgorgement, imprisonment, possible exclusion from participation in federal and state funded healthcare programs, contractual damages, and the curtailment or restricting of our operations, as well as additional reporting obligations and oversight if we become subject to a corporate integrity agreement or other agreement to resolve allegations of non-compliance with these laws. Any action for violation of these laws, even if successfully defended, could cause us to incur significant legal expenses and divert management’s attention from the operation of the business.
Privacy and Data Security
In the ordinary course of business, we and the third parties upon which we rely collect, receive, store, or otherwise process personal data, including information we may collect about participants in our clinical trials. This personal data can also include data that is considered “sensitive” personal data under privacy laws, policies, or agreements to which we are subject. Accordingly, we are, or we may become, subject to numerous, evolving privacy and data security obligations, including global, federal, state, and local laws, regulations, guidance, industry standards, external and internal privacy and security policies, contractual requirements, and other obligations related to privacy and data security.
These privacy and data security laws are evolving and may impose potentially conflicting obligations. Such obligations may include, without limitation, federal health information privacy laws, state information security and data breach notification laws, state health information privacy laws, and federal and state consumer protection laws (e.g., the Federal Trade Commission Act). At the state level, numerous U.S.
states have enacted comprehensive privacy laws that impose certain obligations on covered businesses, including providing specific disclosures in privacy notices and affording individuals certain rights concerning their personal data. Similar laws are being considered in several other states, as well as at the federal and local levels, and we expect more states to pass similar laws in the future. Additionally, a smaller number of states have passed or are considering laws governing the privacy of consumer health data. While existing state consumer privacy laws provide exemptions for data processed in the context of clinical trials, the continued development of complex requirements at the state level may further complicate compliance efforts and may impact our business activities, including our identification of research subjects, relationships with business partners and ultimately the marketing and distribution of our products, and are examples of the increasingly stringent and evolving regulatory frameworks related to personal data processing, as more fully discussed in the section titled “Risk Factors.”
Additionally, to the extent we collect personal data outside of the United States, through clinical trials or otherwise, we are, or may become, subject to foreign data and data security laws, such as the European Union’s General Data Protection Regulation 2016/679 (“EU GDPR”) and other national data protection legislation in force in relevant EEA Member States, and the EU GDPR as it forms part of UK law by virtue of section 3 of the European Union (Withdrawal) Act 2018. Foreign privacy and data security laws impose significant and complex compliance obligations on entities that are subject to those laws and may impose significant sanctions for non-compliance (including fines of up to the greater of 20 million Euros (17.5 million GBP for the UK) or 4% of worldwide annual revenue), as more fully discussed in the section titled “Risk Factors.”
Facilities
Our corporate headquarters is located in Cambridge, Massachusetts, where we lease and occupy approximately 51,726 square feet of office and laboratory space at 399 Binney Street, Cambridge, Massachusetts. The current term of such lease expires in September 2031.
We believe that our facilities are sufficient for our current needs and for the foreseeable future. To meet the future needs of our business, we may lease additional or alternate space. We believe that suitable additional or substitute space at commercially reasonable terms will be available as needed to accommodate any future expansion of our operations.
Employees and Human Capital Resources
As of September 1, 2026, we had 81 full-time employees. Within our workforce, 71 employees were engaged in research and development, and 10 were engaged in business development, finance, legal, and general management and administration as of September 1, 2026. None of our employees are represented by labor unions or covered by collective bargaining agreements. We consider our relationship with our employees to be good.
Our human capital resources objectives include, as applicable, identifying, recruiting, retaining, incentivizing and integrating our existing and new employees, advisors and consultants. The principal purposes of our equity incentive plans are to attract, retain and reward personnel through the granting of stock-based compensation awards in order to increase shareholder value and the success of our company by motivating such individuals to perform to the best of their abilities and achieve our objectives. Our equity incentive plans are heavily weighted towards performance-based vesting (rather than time-based vesting), which further aligns employee incentives to achieve company milestones and drive shareholder value.
We believe that our future success largely depends upon our continued ability to attract and retain highly skilled employees. We provide our employees with competitive salaries and bonuses, opportunities for equity ownership, development programs that enable continued learning and growth, and a robust employment package that promotes well-being across all aspects of their lives, including health care, retirement planning and paid time off.
Legal Proceedings
From time to time, we may become involved in legal proceedings arising from the ordinary course of business. We record a liability for such matters when it is probable that future losses will be incurred and that such losses can be reasonably estimated. Significant judgment by us is required to determine both probability and the estimated amount. Our management is currently not aware of any legal matters that could have a material effect on our financial position, results of operations or cash flows.
MANAGEMENT
Executive Officers and Directors
The following table sets forth information regarding our executive officers and directors as of the date of this prospectus.
|
Name
|
|
|
Age
|
|
|
Position
|
|
|
Executive Officers:
|
|
|
|
|
|
|
|
|
|
|
|
Andrew (Andy) Orth, M.B.A.
|
|
|
|
|
56 |
|
|
|
President, Chief Executive Officer and Director
|
|
|
Keith Regnante, M.B.A.
|
|
|
|
|
57 |
|
|
|
Chief Financial Officer
|
|
|
Baisong Mei, M.D., Ph.D.
|
|
|
|
|
62 |
|
|
|
Chief Medical Officer
|
|
|
Tracy Zimmermann, Ph.D.
|
|
|
|
|
57 |
|
|
|
Chief Scientific Officer
|
|
|
Sebastian Trousil, Ph.D.
|
|
|
|
|
42 |
|
|
|
Chief Operating Officer
|
|
|
Non-Employee Directors:
|
|
|
|
|
|
|
|
|
|
|
|
John Maraganore, Ph.D.
|
|
|
|
|
63 |
|
|
|
Executive Chair
|
|
|
Barry Greene(2)(3)
|
|
|
|
|
63 |
|
|
|
Director
|
|
|
Robert Nelsen, M.B.A.(1)
|
|
|
|
|
63 |
|
|
|
Director
|
|
|
Saraswathy (Sara) V. Nochur, Ph.D.(1)(3)
|
|
|
|
|
66 |
|
|
|
Director
|
|
|
Ron Philip(1)(2)
|
|
|
|
|
52 |
|
|
|
Director
|
|
(1)
Member of the culture and compensation committee.
(2)
Member of the audit committee.
(3)
Member of the nominating and corporate governance committee.
Executive Officers
Andrew (Andy) Orth, M.B.A., has served as our President, Chief Executive Officer and as a member of our board of directors since December 2024. Prior to joining us, Mr. Orth served as Chief Commercial Officer of Krystal Biotech, Inc. (Nasdaq: KRYS), a global, commercial-stage biotechnology company, from May 2021 to August 2023, where he led its global commercial strategy and execution. Previously, Mr. Orth served as Senior Vice President, U.S. Region at Alnylam (Nasdaq: ALNY), a global commercial-stage biopharmaceutical company, from 2016 to 2021, where he contributed to the launch of three genetic medicines for rare diseases and the build-out of Alnylam’s commercial infrastructure in the United States and Europe. Prior to Alnylam, Mr. Orth served as Vice President, Global Commercial Strategy at Biogen Inc. (Nasdaq: BIIB) from 2009 to 2016. Earlier in his career, Mr. Orth held positions in global commercial strategy, commercial operations, corporate strategy and finance at Russell Reynolds Associates Inc., Genzyme Corp. (subsequently acquired by Sanofi S.A. (Nasdaq: SNY) (“Sanofi”)), Amgen Inc. (Nasdaq: AMGN) and the University of California, San Francisco. Mr. Orth received a B.S. in Philosophy and Psychology from the University of Wisconsin and an M.B.A. from Cornell University.
We believe that Mr. Orth is qualified to serve as a member of our board of directors because of his extensive executive leadership and commercial experience across the biopharmaceutical industry, including senior leadership roles at publicly traded biotechnology companies.
Keith Regnante, M.B.A., has served as our Chief Financial Officer since August 2026. Prior to joining us, he served as Chief Financial Officer of Keros Therapeutics, Inc. (Nasdaq: KROS), a clinical-stage biopharmaceutical company, from February 2020 to August 2026. Previously, from August 2016 to January 2020, Mr. Regnante served as Chief Financial Officer of Wave Life Sciences Ltd. (Nasdaq: WVE), a clinical-stage biotechnology company. From February 2014 to August 2016, Mr. Regnante served as Vice President of Finance at Shire plc (formerly Nasdaq: SHPG), a global biotechnology company subsequently acquired by Takeda Pharmaceutical Company Limited, where he was a member of the Financial Leadership Team and the R&D Leadership Team. Earlier in his career, Mr. Regnante served as Head of R&D Finance at ARIAD Pharmaceuticals, Inc. and held
finance leadership positions of increasing responsibility at Biogen Inc., including Senior Director of Corporate Finance and Senior Director of Global R&D Finance. Mr. Regnante began his career as a consultant at The Boston Consulting Group. Mr. Regnante received a B.A. in Economics from Tufts University and an M.B.A. from the MIT Sloan School of Management.
Baisong Mei, M.D., Ph.D., has served as our Chief Medical Officer since March 2025. Previously, Dr. Mei served as Executive Vice President and Chief Medical Officer at Editas Medicine, Inc. (Nasdaq: EDIT) (“Editas”), a gene editing company developing transformative medicines for serious diseases, from July 2022 to December 2024. Prior to Editas, Dr. Mei served as Senior Global Project Head in Rare Disease and Rare Blood Disorders at Sanofi, a global biopharmaceutical company, from January 2017 to July 2022. Before joining Sanofi, Dr. Mei served as Therapeutic Area Head for Hematology Clinical Development at Biogen Inc. Earlier in his career, Dr. Mei held roles in drug discovery research and chemistry, manufacturing and controls development at Biogen and Bayer AG. Dr. Mei received a Bachelor of Medicine from Bengbu Medical University, China, a Master of Medicine from Wuhan University School of Medicine, China, and a Ph.D. in Biochemistry and Molecular Biology from the University of Toledo, and held postdoctoral fellowships at the University of California San Francisco and the University of California Berkeley.
Tracy Zimmermann, Ph.D., has served as our Chief Scientific Officer since May 2024. Prior to joining us, Dr. Zimmermann served in roles of increasing responsibility at Generation Bio Co. (Nasdaq: GBIO) (acquired by XOMA Royalty Corporation (Nasdaq: XOMA)), a biotechnology company, most recently as Chief Development Officer from November 2020 to January 2024, after serving as Vice President, Preclinical Research and Translational Biology from October 2018 to November 2020. Previously, Dr. Zimmermann spent 15 years at Alnylam, serving in research leadership positions spanning RNAi platform development and early-stage therapeutic programs, where she contributed to the development of ONPATTRO, GIVLAARI and AMVUTTRA. Earlier in her career, Dr. Zimmermann served as a Senior Scientist at Variagenics, Inc. (subsequently merged with Hyseq Pharmaceuticals Inc., which was renamed Nuvelo, Inc.). Dr. Zimmermann received a B.S. in Biochemistry from Lehigh University and a Ph.D. in Chemistry from the University of Colorado Boulder and completed a postdoctoral fellowship at Harvard Medical School.
Sebastian Trousil, Ph.D., one of our co-founders, has served as our Chief Operating Officer since September 2023. Prior to joining us, Dr. Trousil served as Principal, New Ventures at RTW Investments, LP, an investment firm, from March 2021 to February 2023. During that time, Dr. Trousil also served as Chief Operating Officer of Yarrow Biotechnology, Inc., a biotechnology company, from January 2021 to February 2023. Dr. Trousil co-founded Harbinger Health, Inc., a liquid biopsy company focused on early cancer detection, and served in leadership roles during its early development. Earlier in his career, Dr. Trousil worked at Flagship Pioneering, a life sciences venture capital and company-building firm. Dr. Trousil received an M.Pharm. from the University of Vienna, Austria, and a Ph.D. in Pharmacology and Cancer Research from Imperial College London, England and completed postdoctoral research at Harvard Medical School and Massachusetts General Hospital, where he was awarded the Tosteson & Fund for Medical Discovery Postdoctoral Fellowship.
Non-Executive Directors
John Maraganore, Ph.D., one of our co-founders, has served as our Executive Chair and a member of our board of directors since September 2023. Dr. Maraganore has served as Principal of JMM Innovations, LLC, a strategic consulting firm, since January 2022, and as Co-Chief Executive Officer of Corsera Health, Inc., a privately held biotechnology company, since September 2024. He also currently serves as a Venture Partner at ARCH Venture Partners, a Venture Advisor at Atlas Venture, a Senior Advisor at Blackstone Life Sciences, a Senior Advisor at Jefferies Financial Services, and an Executive Partner at RTW Investments, LP, each an investment and venture capital firm. Previously, Dr. Maraganore served as founding Chief Executive Officer and as a member of the board of directors of Alnylam from December 2002 to December 2021, and as President of Alnylam from December 2002 to December 2007. Dr. Maraganore currently serves on the boards of directors of Kymera Therapeutics, Inc. (Nasdaq: KYMR), Beam Therapeutics Inc. (Nasdaq: BEAM) (“Beam Therapeutics”), Rapport Therapeutics, Inc. (Nasdaq: RAPP), Hemab Therapeutics Holdings, Inc. (Nasdaq: COAG) and Takeda
Pharmaceutical Company Limited (NYSE: TAK). He previously served on the boards of directors of Agios Pharmaceuticals, Inc. (Nasdaq: AGIO) from June 2010 to May 2023 and ProKidney Corp. (Nasdaq: PROK) from August 2022 to May 2024. Dr. Maraganore received a B.A. in Biological Sciences and M.S. and Ph.D. degrees in Biochemistry and Molecular Biology, each from the University of Chicago.
We believe that Dr. Maraganore is qualified to serve as our Executive Chair and a member of our board of directors because of his extensive executive leadership experience in the biopharmaceutical industry, including his service as Chief Executive Officer of Alnylam and his service on the boards of directors of multiple publicly traded biopharmaceutical companies.
Barry Greene has served as a member of our board of directors since September 2024. Mr. Greene served as President and Chief Executive Officer of Sage Therapeutics, Inc. (Nasdaq: SAGE) (“Sage”), a biopharmaceutical company, from December 2020 until it was acquired by Supernus Pharmaceuticals, Inc. (Nasdaq: SUPN) in July 2025. Prior to joining Sage, Mr. Greene served as President of Alnylam from 2007 through September 2020 and as Chief Operating Officer from September 2003 to December 2020. Prior to joining Alnylam, Mr. Greene served as General Manager of Oncology at Millennium Pharmaceuticals, Inc. Earlier in his career, Mr. Greene served as Executive Vice President and Chief Business Officer at Mediconsult.com, Inc., Vice President of Marketing and Customer Services and Vice President, Strategic Integration at AstraZeneca PLC (Nasdaq: AZN), and Partner at Andersen Consulting. Mr. Greene currently serves on the board of directors of Karyopharm Therapeutics, Inc. (Nasdaq: KPTI) and Dyne Therapeutics, Inc. (Nasdaq: DYN). He previously served on the boards of directors of Acorda Therapeutics, Inc. (Nasdaq: ACOR) from January 2007 to August 2021 and BCLS Acquisition Corporation (Nasdaq: BLSA) from October 2020 to October 2022. Mr. Greene received a B.S. in Industrial Engineering from the University of Pittsburgh and completed the Senior Scholars executive program at Duke University’s Medical School and Fuqua School of Business.
We believe that Mr. Greene’s extensive operational and executive leadership experience across biopharmaceutical companies, including his tenure as a public company chief executive officer and his significant commercial and business development expertise, qualify him to serve as a member of our board of directors.
Robert Nelsen, M.B.A., has served as a member of our board of directors since July 2023. Mr. Nelsen co-founded ARCH Venture Partners, a venture capital firm focused on early-stage technology companies, in 1986, and has served as a Managing Director of ARCH Venture Partners or its affiliated entities since 1994. Mr. Nelsen currently serves on the boards of directors of several publicly-traded biotechnology companies, including Prime Medicine, Inc. (Nasdaq: PRME), Sana Biotechnology, Inc. (Nasdaq: SANA), and Hua Medicine, Inc. (HKEX: 2552). Mr. Nelsen also currently serves on the boards of directors of a number of privately-held biotechnology companies and previously served on the boards of directors of several publicly-traded biotechnology and biopharmaceutical companies, including VIR Biotechnology, Inc. (Nasdaq: VIR) from 2017 to May 2025, Lyell Immunopharma, Inc. (Nasdaq: LYEL) from 2019 to May 2025, Seaport Therapeutics, Inc. (Nasdaq: SPTX) from 2024 to April 2026, Brii Biosciences Limited (HKEX: 2137) from 2019 to July 2024, Neumora Therapeutics, Inc. (Nasdaq: NMRA) from 2020 to September 2023, Revolution Healthcare Acquisition Corp. (Nasdaq: REVH) from 2021 to April 2022, Denali Therapeutics Inc. (Nasdaq: DNLI) from 2015 to June 2022, Beam Therapeutics from 2017 to June 2021, and Karuna Therapeutics, Inc. (Nasdaq: KRTX) (subsequently acquired by Bristol Myers Squibb) from 2018 to June 2021. Mr. Nelsen received his B.S. from the University of Puget Sound in Economics and Biology and his M.B.A. from the University of Chicago.
We believe that Mr. Nelsen is qualified to serve on our board of directors because of his extensive experience as a venture capitalist, building and serving on the boards of many public and private emerging companies, including multiple life sciences, biotechnology and pharmaceutical companies.
Saraswathy (Sara) V. Nochur, Ph.D., has served as a member of our board of directors since February 2025. Dr. Nochur has served as an independent consultant in regulatory affairs and drug development since June 2024. Previously, Dr. Nochur served in roles of increasing responsibility at Alnylam, most recently as Chief Diversity, Equity and Inclusion Officer from January 2021 to May 2024. Prior to that, she oversaw global regulatory affairs at Alnylam as Vice President of Regulatory Affairs,
Senior Vice President of Regulatory Affairs, and Chief Regulatory Officer from May 2005 to December 2020, and she also served as a member of Alnylam’s management board. Prior to joining Alnylam, Dr. Nochur served in roles of increasing responsibility in regulatory affairs at The Medicines Company, most recently as Vice President of Regulatory Affairs. Dr. Nochur previously served on the boards of directors of Decibel Therapeutics, Inc. (Nasdaq: DBTX) from December 2021 until its acquisition by Regeneron Pharmaceuticals, Inc. (Nasdaq: REGN) in September 2023 and Marinus Pharmaceuticals, Inc. (Nasdaq: MRNS) from March 2021 to November 2024. Dr. Nochur received a B.S. in Microbiology and Chemistry, an M.S. in Microbiology from the University of Bombay, India, and a Ph.D. in Biochemical Engineering from the Massachusetts Institute of Technology.
We believe that Dr. Nochur is qualified to serve on our board of directors due to her extensive experience in drug development, regulatory strategy and the regulatory approval process.
Ron Philip has served as a member of our board of directors since July 2024. Mr. Philip served as Chief Executive Officer and as a member of the board of directors of Orbital Therapeutics, Inc. (“Orbital”), a biopharmaceutical company focused on RNA-based medicines, from August 2024 until its acquisition by Bristol Myers Squibb (NYSE: BMY) in December 2025. Prior to joining Orbital, Mr. Philip served in roles of increasing responsibility at Spark Therapeutics, Inc. (formerly Nasdaq: ONCE) from May 2017 to August 2024, most recently as Chief Executive Officer. Prior to joining Spark Therapeutics, Mr. Philip served in various roles at Pfizer Inc. (NYSE: PFE) from November 2009 to October 2016. Mr. Philip currently serves on the board of directors of Harmony Biosciences Holdings, Inc. (Nasdaq: HRMY), Avalo Therapeutics, Inc. (Nasdaq: AVTX) and serves as Executive Chair of CREATE Medicines, Inc. Mr. Philip received a B.S. in Computer Information Systems from Drexel University.
We believe that Mr. Philip is qualified to serve on our board of directors because of his extensive experience in the life sciences industry, including leading teams focused on the development and commercialization of novel treatments, and his demonstrated business and strategic acumen.
Family Relationships
There are no family relationships among any of our executive officers or directors.
Composition of Our Board of Directors
Our business and affairs are managed under the direction of our board of directors, which currently consists of six members. The primary responsibilities of our board of directors are to provide oversight, strategic guidance, counseling and direction to our management. Our board of directors meets on a regular basis and additionally as required.
Certain members of our board of directors were elected under the provisions of our amended and restated certificate of incorporation and agreements with our stockholders. These board composition provisions will terminate upon the completion of this offering. Upon the termination of these provisions, there will be no further contractual obligations regarding the election of our directors. Our nominating and corporate governance committee and our board of directors may therefore consider a broad range of factors relating to the qualifications and background of nominees. Our nominating and corporate governance committee’s and our board of directors’ priority in selecting board members is the identification of persons who will further the interests of our stockholders through their established record of professional accomplishment, the ability to contribute positively to the collaborative culture among board members, knowledge of our business, understanding of the competitive landscape, professional and personal experiences, and expertise relevant to our growth strategy. Our directors hold office until their successors have been elected and qualified or until the earlier of their resignation or removal. Our amended and restated certificate of incorporation, which will become effective immediately prior to the completion of this offering, and our amended and restated bylaws, which will become effective upon the effectiveness of the registration statement of which this prospectus forms a part, will also provide that our directors may be removed only for cause by the affirmative vote of the holders of at least two-thirds of the votes that all our stockholders would be entitled to cast in an annual election of directors, and that any vacancy on our board of directors, including a vacancy resulting from an enlargement of our board of directors, may be filled only by vote of a majority of our directors then in office.
Staggered Board
Our amended and restated certificate of incorporation, which will become effective immediately prior to the completion of this offering, and our amended and restated bylaws, which will be effective upon the effectiveness of the registration statement of which this prospectus forms a part, will permit our board of directors to establish the authorized number of directors from time to time by resolution. Each director serves until the expiration of the term for which such director was elected or appointed, or until such director’s earlier death, resignation or removal. In accordance with our amended and restated certificate of incorporation, our board of directors will be divided into three classes with staggered three-year terms. At each annual meeting of stockholders, the successors to directors whose terms then expire will be elected to serve from the time of election and qualification until the third annual meeting following election. Our directors will be divided among the three classes as follows:
•
the Class I directors will be Andrew Orth, M.B.A. and Ron Philip, and their terms will expire at our first annual meeting of stockholders following this offering, to be held in 2027;
•
the Class II directors will be John Maraganore, Ph.D. and Barry Greene, and their terms will expire at our second annual meeting of stockholders following this offering, to be held in 2028; and
•
the Class III directors will be Robert Nelsen, M.B.A. and Sara Nochur, Ph.D., and their terms will expire at our third annual meeting of stockholders following this offering, to be held in 2029.
We expect that any additional directorships resulting from an increase in the number of directors will be distributed among the three classes so that, as nearly as possible, each class will consist of one third of the directors. The division of our board of directors into three classes with staggered three-year terms may delay or prevent a change of our management or a change in control.
This classification of our board of directors may have the effect of delaying or preventing changes in control of our company.
Director Independence
Under the listing standards, requirements and rules of The Nasdaq Stock Market LLC (the “Nasdaq Listing Rules”), independent directors must comprise a majority of our board of directors as a listed company within one year of the listing date. In addition, the Nasdaq Listing Rules require that, subject to specified exceptions, and phase-in periods following its initial public offering, each member of a listed company’s audit committee must have at least three members and its compensation committee must have two members, and each member of its audit, compensation, and nominating and governance committees be independent within twelve months from the date of listing. Audit committee members must also satisfy additional independence criteria, including those set forth in Rule 10A-3 under the Exchange Act, and compensation committee members must also satisfy the independence criteria set forth in Rule 10C-1 under the Exchange Act. Under the Nasdaq Listing Rules, a director will only qualify as an “independent director” if, in the opinion of that company’s board of directors, that person does not have a relationship that would interfere with the exercise of independent judgment in carrying out the responsibilities of a director. In order to be considered independent for purposes of Rule 10A-3 under the Exchange Act, a member of an audit committee of a listed company may not, other than in his or her capacity as a member of the audit committee, the board of directors, or any other board committee: (i) accept, directly or indirectly, any consulting, advisory, or other compensatory fee from the listed company or any of its subsidiaries, other than compensation for board service; or (ii) be an affiliated person of the listed company or any of its subsidiaries. In order to be considered independent for purposes of Rule 10C-1 under the Exchange Act, the board of directors must consider, for each member of a compensation committee of a listed company, all factors specifically relevant to determining whether a director has a relationship to such company which is material to that director’s ability to be independent from management in connection with the duties of a compensation committee member, including, but not limited to: the source of compensation of the director, including any consulting advisory or other compensatory fee paid by such company to the director, and whether the director is affiliated with the company or any of its subsidiaries or affiliates.
Our board of directors has undertaken a review of the independence of each director. Based on information provided by each director concerning her or his background, employment, and affiliations, including family relationships, our board of directors has determined that each of Barry Greene, Robert Nelsen, M.B.A., Sara Nochur, Ph.D. and Ron Philip does not have relationships that would interfere with the exercise of independent judgment in carrying out the responsibilities of a director and that each of these directors is “independent” as that term is defined under the Nasdaq Listing Rules. Our board of directors has determined that Mr. Orth, by virtue of his position as our current President and Chief Executive Officer, and Dr. Maraganore, by virtue of his current consulting arrangement with the Company, are not independent under applicable rules and regulations of the SEC and the Nasdaq Listing Rules. In making these determinations, our board of directors considered the current and prior relationships that each non-employee director has with our company and all other facts and circumstances our board of directors deemed relevant in determining their independence, including the beneficial ownership of our shares by each non-employee director and the transactions described in the section titled “Certain Relationships and Related Person Transactions.”
Board Policies
In connection with this offering, we have adopted policies and procedures for director candidates for our nominating and corporate governance committee, which will provide that factors, such as a candidate’s character, judgment, skills, education, expertise, and absence of conflicts of interest should be considered in determining director candidates. Our priority in selection of board members will be identification of members who will further the interests of our stockholders through their established records of professional accomplishment, their ability to contribute positively to the collaborative culture among board members, and their knowledge of our business and understanding of the competitive landscape in which we operate and adherence to high ethical standards.
Board Leadership Structure and Board’s Role in Risk Oversight
Currently, the role of executive chair of our board of directors is separated from the role of Chief Executive Officer. We believe that separating these positions allows our Chief Executive Officer to focus on our day-to-day business, while allowing the executive chair of our board to lead the board of directors in its fundamental role of providing advice to and independent oversight of management. Our board of directors recognizes the time, effort and energy that the chief executive officer is required to devote to his position in the current business environment, as well as the commitment required to serve as our chair of our board of directors, particularly as the board of directors’ oversight responsibilities continue to grow. While our amended and restated bylaws and corporate governance guidelines will not require that our board chair and chief executive officer positions be separate, our board of directors believes that having separate positions is the appropriate leadership structure for us at this time and demonstrates our commitment to good corporate governance. Our board of directors will adopt, effective prior to the completion of this offering, corporate governance guidelines that will provide that the board of directors may appoint a lead independent director. The lead independent director will be responsible for calling and presiding over separate meetings of the independent directors. The lead independent director will preside over periodic meetings of independent directors, serve as a liaison between the chair and the independent directors and perform such additional duties as the board of directors may otherwise determine and delegate.
Risk is inherent with every business, and how well a business manages risk can ultimately determine its success. We face a number of risks, including risks relating to our financial condition, development, and commercialization activities, operations, strategic direction, and intellectual property as more fully discussed in the section titled “Risk Factors.” Management is responsible for the day-to-day management of risks we face, while our board of directors, as a whole and through its committees, has responsibility for the oversight of risk management. In its risk oversight role, our board of directors has the responsibility to satisfy itself that the risk management processes designed and implemented by management are adequate and functioning as designed.
The role of the board of directors in overseeing the management of our risks is conducted primarily through committees of the board of directors, as disclosed in the descriptions of each of the committees
below and in the charters of each of the committees. The full board of directors (or the appropriate board committee in the case of risks that are under the purview of a particular committee) discusses with management our major risk exposures, their potential impact on us, and the steps we take to manage them. When a board committee is responsible for evaluating and overseeing the management of a particular risk or risks, the chairperson of the relevant committee reports on the discussion to the full board of directors during the committee reports portion of the next board meeting. This enables the board of directors and its committees to coordinate the risk oversight role, particularly with respect to risk interrelationships.
Committees of Our Board of Directors
Our board of directors has established an audit committee, a culture and compensation committee, and a nominating and corporate governance committee. The composition and responsibilities of each of the committees of our board of directors are described below. Members serve on these committees until their resignation or until otherwise determined by our board of directors. Each committee has adopted a written charter that satisfies the applicable rules and regulation of the SEC and the Nasdaq Listing Rules, which we will post to our website at www.citytx.com upon the completion of this offering. Information contained on, or accessible through, our website is not a part of this prospectus, and the inclusion of our website address in this prospectus is only an inactive textual reference. Our board of directors may establish other committees as it deems necessary or appropriate from time to time.
Audit Committee
Upon the effectiveness of the registration statement of which this prospectus forms a part, our audit committee will consist of Barry Greene and Ron Philip, and the chair of our audit committee will be Barry Greene. Our board of directors has determined that each member of the audit committee is independent under Nasdaq Listing Rules and Rule 10A-3(b)(1) of the Exchange Act and can read and understand fundamental financial statements in accordance with applicable requirements. We intend to rely on the phase-in provisions of Nasdaq Listing Rule 5615, which permit companies listing in connection with their initial public offering to phase-in compliance with the heightened audit committee independence requirements. Specifically, we will be required to have an audit committee of at least three directors that satisfy the heightened audit committee independence requirements within one year of listing. We intend to comply with such requirements within the allotted timeframe. Our board of directors has also determined that Barry Greene is an “audit committee financial expert” within the meaning of SEC regulations. In arriving at these determinations, our board of directors has examined each audit committee member’s scope of experience and the nature of their employment in the corporate finance sector.
The primary purpose of the audit committee will be to discharge the responsibilities of our board of directors with respect to our corporate accounting and financial reporting processes, systems of internal control and financial-statement audits, and to oversee our independent registered public accounting firm. Specific responsibilities of our audit committee will include:
•
helping our board of directors oversee our corporate accounting and financial reporting processes;
•
coordinating the oversight and reviewing the adequacy of our internal control over financial reporting;
•
managing the selection, engagement, qualifications, independence, and performance of a qualified firm to serve as the independent registered public accounting firm to audit our financial statements;
•
discussing the scope and results of the audit with the independent registered public accounting firm, and reviewing, with management and the independent accountants, our interim and year-end operating results;
•
developing procedures for employees to submit concerns anonymously about questionable accounting or audit matters;
•
reviewing and approving related person transactions;
•
reviewing the overall audit plan with our independent registered public accounting firm and members of management responsible for preparing our financial statements;
•
recommending, based upon the audit committee’s review and discussions with management and our independent registered public accounting firm, whether our audited financial statements shall be included in our Annual Report on Form 10-K;
•
overseeing the preparation of our annual proxy statement, reviewing with management our financial statements to be included in our quarterly reports to be filed with the SEC, and reviewing with management the “Management’s Discussion and Analysis of Financial Condition and Results of Operations” disclosures in our periodic reports filed with the SEC;
•
monitoring the integrity of our financial statements and our compliance with legal and regulatory requirements as they relate to our financial statements and accounting matters;
•
overseeing our risk management policies, procedures and practices, including those related to cybersecurity; and
•
approving, or, as permitted, pre-approving, audit and permissible non-audit services to be performed by the independent registered public accounting firm.
Our audit committee will operate under a written charter, which will be effective upon the effectiveness of the registration statement of which this prospectus forms a part, that satisfies the applicable Nasdaq Listing Rules.
Culture and Compensation Committee
Upon the effectiveness of the registration statement of which this prospectus forms a part, our culture and compensation committee will consist of Ron Philip, Sara Nochur, Ph.D. and Robert Nelsen, M.B.A., and the chair of our culture and compensation committee will be Ron Philip. Our board of directors has determined that each member of the culture and compensation committee is independent under the Nasdaq Listing Rules and is a “non-employee director” as defined in Rule 16b-3 promulgated under the Exchange Act.
The primary purpose of our culture and compensation committee will be to discharge the responsibilities of our board of directors in overseeing our compensation policies, plans, and programs and to review and determine the compensation to be paid to our executive officers, directors, and other senior management, as appropriate. Specific responsibilities of our culture and compensation committee will include:
•
annually reviewing and recommending to our board of directors the corporate goals and objectives relevant to the compensation of our Chief Executive Officer;
•
evaluating the performance of our Chief Executive Officer in light of such corporate goals and objectives and, based on such evaluation, recommending to our board of directors the compensation of our Chief Executive Officer;
•
reviewing and approving the compensation arrangements with our other executive officers and certain other senior management;
•
reviewing and recommending to our board of directors the compensation paid to our directors;
•
administering our equity incentive plans and other benefit programs;
•
overseeing and administering our compensation and similar plans;
•
reviewing and approving our policies and procedures for the grant of equity-based awards;
•
preparing our culture and compensation committee report if and when required by SEC rules;
•
reviewing and establishing general policies relating to compensation and benefits of our employees, including our overall compensation philosophy;
•
reviewing and discussing annually with management our “Compensation Discussion and Analysis,” if and when required, to be included in our annual proxy statement; and
•
assisting our board of directors in overseeing our strategies, programs and initiatives related to employee development, engagement, pay equity, succession, and workplace culture and inclusion, and periodically reviewing and making recommendations regarding succession planning for our Chief Executive Officer and other key officers.
Our culture and compensation committee will operate under a written charter, which will be effective upon the effectiveness of the registration statement of which this prospectus forms a part, that satisfies the applicable Nasdaq Listing Rules.
Nominating and Corporate Governance Committee
Upon the effectiveness of the registration statement of which this prospectus forms a part, our nominating and corporate governance committee will consist of Barry Greene and Saraswathy (Sara) V. Nochur, Ph.D., and the chair of our nominating and corporate governance committee will be Barry Greene. Our board of directors has determined that each member of the nominating and corporate governance committee is independent under the Nasdaq Listing Rules, a non-employee director, and free from any relationship that would interfere with the exercise of his or her independent judgment.
The primary purpose of the nominating and corporate governance committee is to discharge the responsibilities of our board of directors with respect to our corporate governance functions and to identify, communicate with, evaluate and recommend candidates for our board of directors. Specific responsibilities of our nominating and corporate governance committee include:
•
identifying and evaluating candidates, including the nomination of incumbent directors for reelection and nominees recommended by stockholders, to serve on our board of directors;
•
considering and making recommendations to our board of directors regarding the composition and chairmanship of the committees of our board of directors;
•
instituting plans or programs for the continuing education of our board of directors and orientation of new directors;
•
developing and recommending to the board of directors criteria for board and committee membership;
•
establishing procedures for identifying and evaluating board of director candidates, including nominees recommended by stockholders;
•
developing and making recommendations to our board of directors regarding corporate governance guidelines and matters and periodically reviewing and reassessing the adequacy of our corporate governance guidelines and Code of Conduct and Business Ethics; and
•
overseeing annual evaluations of the board of directors’ performance, including committees of the board of directors and management.
Our nominating and corporate governance committee will operate under a written charter, which will be effective upon the effectiveness of the registration statement of which this prospectus forms a part, that satisfies the applicable Nasdaq Listing Rules.
Code of Business Conduct and Ethics
In connection with this offering, we have adopted a written code of business conduct and ethics that applies to all our employees, officers and directors, effective upon the effectiveness of the registration statement of which this prospectus forms a part. This includes our principal executive officer, principal financial officer and principal accounting officer or controller, or persons performing similar functions. The full text of our code of business conduct and ethics will be posted on our website at www.citytx.com. We intend to disclose on our website any future amendments of our code of business
conduct and ethics or waivers that exempt any principal executive officer, principal financial officer, principal accounting officer or controller, persons performing similar functions or our directors from provisions in the code of business conduct and ethics. Information contained on, or accessible through, our website is not a part of this prospectus, and the inclusion of our website address in this prospectus is only an inactive textual reference.
Compensation Committee Interlocks and Insider Participation
None of the members of the culture and compensation committee is currently, or has been at any time, one of our officers or employees. None of our officers currently serves, or has served during the last calendar year, as a member of the board of directors or culture and compensation committee of any entity that has one or more executive officers serving as a member of our board of directors or culture and compensation committee.
Compensation Recovery
In accordance with the requirements of the SEC and Nasdaq Listing Rules, our board of directors has adopted a compensation recovery policy, which will become effective upon the effectiveness of the registration statement of which this prospectus is part. The compensation recovery policy will provide that in the event we are required to prepare a restatement of financial statements due to material noncompliance with any financial reporting requirement under securities laws, we will seek to recover any incentive-based compensation that was based upon the attainment of a financial reporting measure and that was received by any current or former executive officer during the three-year period preceding the date that the restatement was required if such compensation exceeds the amount that the executive officers would have received based on the restated financial statements.
Limitations on Liability and Indemnification
As permitted by Delaware law, provisions in our amended and restated certificate of incorporation, which will become effective immediately prior to the completion of this offering, and our amended and restated bylaws, which will be effective upon the effectiveness of the registration statement of which this prospectus forms a part, limit or eliminate the personal liability of directors and officers for a breach of their fiduciary duty of care as a director or officer. The duty of care generally requires that, when acting on behalf of the corporation, a director or officer exercise an informed business judgment based on all material information reasonably available to him or her. Consequently, a director or officer will not be personally liable to us or our stockholders for monetary damages or breach of fiduciary duty as a director or officer, except for liability for:
•
any breach of the officer or director’s duty of loyalty to us or our stockholders;
•
any act or omission not in good faith or that involves intentional misconduct or a knowing violation of law;
•
for our directors, unlawful payments of dividends or unlawful stock repurchases, or redemptions as provided in Section 174 of the DGCL;
•
for our officers, any derivative action by or in the right of the corporation;
•
any act related to unlawful stock repurchases, redemptions or other distributions or payments of dividends; or
•
any transaction from which the director or officer derived an improper personal benefit.
These limitations of liability do not limit or eliminate our rights or any stockholder’s rights to seek non-monetary relief, such as injunctive relief or rescission. These provisions will not alter a director or officer’s liability under other laws, such as the federal securities laws or other state or federal laws. Our amended and restated certificate of incorporation that will become effective upon the completion of this offering also will authorize us to indemnify our officers, directors, and other agents to the fullest extent permitted under Delaware law.
As permitted by Delaware law, our amended and restated bylaws, which will be effective upon effectiveness of the registration statement of which this prospectus forms a part, will provide that:
•
we will indemnify our directors, officers, employees and other agents to the fullest extent permitted by law;
•
we must advance expenses to our directors and officers, and may advance expenses to our employees and other agents, in connection with a legal proceeding to the fullest extent permitted by law; and
•
the rights provided in our amended and restated bylaws are not exclusive.
If Delaware law is amended to authorize corporate action further eliminating or limiting the personal liability of a director or officer, then the liability of our directors or officers will be so eliminated or limited to the fullest extent permitted by Delaware law, as so amended. Our amended and restated bylaws will also permit us to secure insurance on behalf of any officer, director, employee, or other agent for any liability arising out of his or her actions in connection with their services to us, regardless of whether our amended and restated bylaws permit such indemnification. We intend to obtain such insurance.
In addition to the indemnification that will be provided for in our amended and restated certificate of incorporation and amended and restated bylaws, we plan to enter into separate indemnification agreements with each of our directors and executive officers, which may be broader than the specific indemnification provisions contained in the DGCL. These indemnification agreements may require us, among other things, to indemnify our directors and executive officers for some expenses, including attorneys’ fees, expenses, judgments, fines, and settlement amounts incurred by a director or executive officer in any action or proceeding arising out of his service as one of our directors or executive officers or any other company or enterprise to which the person provides services at our request. We believe that these provisions and agreements are necessary to attract and retain qualified individuals to serve as directors and executive officers.
This description of the indemnification provisions of our amended and restated certificate of incorporation, our amended and restated bylaws and our indemnification agreements is qualified in its entirety by reference to these documents, each of which is attached as an exhibit to the registration statement of which this prospectus forms a part.
Insofar as indemnification for liabilities arising under the Securities Act may be permitted to our directors, officers and controlling persons pursuant to the foregoing provisions, or otherwise, we have been advised that, in the opinion of the SEC, such indemnification is against public policy as expressed in the Securities Act, and is, therefore, unenforceable.
There is no pending litigation or proceeding naming any of our directors or officers as to which indemnification is being sought, nor are we aware of any pending or threatened litigation that may result in claims for indemnification by any director or officer.
Rule 10b5-1 Sales Plans
Our directors and executive officers may adopt written plans, known as Rule 10b5-1 plans, in which they will contract with a broker to buy or sell shares of our common stock on a periodic basis. Under a Rule 10b5-1 plan, a broker executes trades pursuant to parameters established by the director or officer when entering into the plan, without further direction from them. The director or officer may amend a Rule 10b5-1 plan in some circumstances and may terminate a plan at any time. Our directors and executive officers also may buy or sell additional shares outside of a Rule 10b5-1 plan when they are not in possession of material nonpublic information subject to compliance with the terms of our insider trading policy. Prior to 180 days after the date of this offering, subject to early termination, the sale of any shares under such plans would be prohibited by the lock-up agreement that the director or officer has entered into with the underwriters.
EXECUTIVE COMPENSATION
The following discussion contains forward-looking statements that are based on our current plans, considerations, expectations, and determinations regarding future compensation programs. The actual amount and form of compensation and the compensation policies and practices that we adopt in the future may differ materially from the programs as summarized in this discussion.
As an emerging growth company and a smaller reporting company, we have opted to comply with the executive compensation disclosure rules applicable to “emerging growth companies” and “smaller reporting companies,” as such terms are defined in the rules promulgated under the Securities Act. The compensation provided to our named executive officers for the fiscal year ended December 31, 2025 (“Fiscal Year 2025”) is detailed in the 2025 Summary Compensation Table and accompanying footnotes and narrative that follow.
Our named executive officers for Fiscal Year 2025 are:
•
Andrew (Andy) Orth, M.B.A., our President and Chief Executive Officer;
•
Baisong Mei, M.D., Ph.D., our Chief Medical Officer; and
•
Tracy Zimmermann, Ph.D., our Chief Scientific Officer.
To date, the compensation of our named executive officers has consisted of a combination of base salary, cash incentive compensation, and long-term incentive compensation, as more fully described below. Our named executive officers, like all full-time employees, are eligible to participate in our health, welfare, and retirement benefit plans. As we transition from a private company to a publicly traded company, we intend to evaluate our compensation values and philosophy and compensation plans and arrangements as circumstances require.
2025 Summary Compensation Table
The following table shows the total compensation earned by, or paid to, our named executive officers for services rendered to us in all capacities during Fiscal Year 2025.
|
Name and Principal Position
|
|
|
Year
|
|
|
Salary
($)
|
|
|
Bonus
($)
|
|
|
Option
Awards(1)
($)
|
|
|
Non-Equity
Incentive Plan
Compensation(2)
($)
|
|
|
All Other
Compensation(3)
($)
|
|
|
Total
($)
|
|
|
Andrew Orth, M.B.A.
President and Chief
Executive Officer
|
|
|
|
|
2025 |
|
|
|
|
|
475,000 |
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
|
|
188,100 |
|
|
|
|
|
14,000 |
|
|
|
|
|
677,100 |
|
|
|
Baisong Mei, M.D., Ph.D.
Chief Medical Officer(4)
|
|
|
|
|
2025 |
|
|
|
|
|
358,077 |
|
|
|
|
|
50,000(5) |
|
|
|
|
|
364,236 |
|
|
|
|
|
110,226 |
|
|
|
|
|
9,500 |
|
|
|
|
|
892,039 |
|
|
|
Tracy Zimmermann, Ph.D.
Chief Scientific Officer
|
|
|
|
|
2025 |
|
|
|
|
|
499,200 |
|
|
|
|
|
— |
|
|
|
|
|
19,831 |
|
|
|
|
|
153,754 |
|
|
|
|
|
14,000 |
|
|
|
|
|
686,785 |
|
|
(1)
The amounts reported represent the aggregate grant date fair value of stock option awards granted in Fiscal Year 2025, computed in accordance with Financial Accounting Standards Board Accounting Standards Codification Topic 718 (“FASB ASC Topic 718”). Such grant date fair values do not take into account any estimated forfeitures related to service-based vesting. The assumptions used in calculating the grant date fair values of the option awards reported in this column are set forth in Note 12 of our consolidated financial statements for Fiscal Year 2025, included elsewhere in this prospectus. The amounts reported in this column reflect the accounting cost for these option awards and do not correspond to the actual economic value that may be received by our named executive officers upon the exercise of the option awards or any sale of the underlying securities.
(2)
The amounts reported represent the annual cash incentive bonuses earned for Fiscal Year 2025 based on achievement of corporate performance measures and individual performance, as described in more detail under the heading “2025 Cash Bonuses” below. The bonus paid to Dr. Mei was prorated to reflect his partial year of employment.
(3)
The amounts reported represent 401(k) employer matching contributions made by us on behalf of each named executive officer under our 401(k) plan.
(4)
Dr. Mei commenced employment with us in March 2025. The amount reported in the salary column represents the amount actually earned for his partial year of employment.
(5)
The amount reported represents a sign-on bonus in the amount of $50,000 in connection with the commencement of Dr. Mei’s employment with us in 2025.
Narrative Disclosure to the 2025 Summary Compensation Table
2025 Base Salaries
Our named executive officers each receive a base salary to compensate them for services rendered to us. Base salaries are intended to provide a fixed component of compensation reflecting the executive’s skill set, experience, role, and responsibilities. Base salaries are expected to be reviewed annually, typically in connection with our annual performance review process, approved by our board of directors or the culture and compensation committee of the board of directors, and may be adjusted from time to time to realign salaries with market levels after taking into account individual responsibilities, performance, and experience.
For Fiscal Year 2025, the base salaries for Mr. Orth, Dr. Mei, and Dr. Zimmermann were $475,000, $475,000, and $499,200, respectively.
In the event that the Company successfully consummates this offering on or prior to December 31, 2026, the annual base salaries for Mr. Orth and Dr. Mei will increase to $600,000 and $515,000, respectively, in each case effective as of the consummation of this offering.
2025 Cash Bonuses
Our named executive officers are eligible to receive annual performance-based cash bonuses, which are designed to provide appropriate incentives to our executives to achieve corporate performance goals and to reward our executives for individual achievement towards these goals. For Fiscal Year 2025, each of the named executive officers was eligible to earn an annual performance bonus based on the achievement of certain corporate performance and individual performance goals. For Fiscal Year 2025, the target annual bonus for Mr. Orth, Dr. Mei, and Dr. Zimmermann was 45%, 35%, and 35% of base salary, respectively. Based on the level of achievement of the applicable corporate performance measures and individual performance goals, annual bonuses for Fiscal Year 2025 were paid at 88% of target. Dr. Mei’s Fiscal Year 2025 annual bonus was prorated based on his start date.
In the event that the Company successfully consummates this offering on or prior to December 31, 2026, the target annual bonuses for Mr. Orth, Dr. Mei, and Dr. Zimmermann will increase to 55%, 40%, and 40% of base salary, respectively, in each case effective as of the consummation of this offering.
Equity Incentive Compensation
Our executive compensation philosophy is designed to align compensation with long-term value creation and the interests of our stockholders. Consistent with this philosophy, we provide annual equity grants to our executive officers, with a significant portion of such grants weighted toward performance-based vesting. We believe equity grants provide our executives with a strong link to our long-term performance, create an ownership culture, and help to align the interests of our executives and our stockholders. In addition, we believe equity grants promote executive retention because they incentivize our executive officers to remain in our service during the vesting period.
During Fiscal Year 2025, we granted Dr. Mei and Dr. Zimmermann stock options to purchase our common stock under the 2023 Plan. Mr. Orth did not receive any equity awards in Fiscal Year 2025. For additional information regarding outstanding equity awards held by our named executive officers as of December 31, 2025, see the “Outstanding Equity Awards at 2025 Fiscal Year-End” table below.
401(k) Plan and Health and Welfare Benefits
During Fiscal Year 2025, we participated in the ADP TotalSource Retirement Savings Plan (the “ADP 401(k) Plan”), a 401(k) plan maintained by ADP TotalSource Group, Inc., our professional employer organization, in which our employees, including our named executive officers, were eligible to participate if they satisfy certain eligibility requirements. Our named executive officers participated in
the ADP 401(k) Plan on the same terms as our other full-time employees. During Fiscal Year 2025, we provided employer contributions under the ADP 401(k) Plan of 4% of each eligible employee’s annual eligible compensation, subject to the safe harbor limit. Beginning in January 1, 2026, we sponsor and maintain our own 401(k) plan through Fidelity for our employees, including our named executive officers. We believe that providing a vehicle for tax-deferred retirement savings through the 401(k) plan adds to the overall desirability of our executive compensation package and further incentivizes our employees, including our named executive officers, in accordance with our compensation policies. Other than the 401(k) plan, we do not provide any qualified or non-qualified retirement or deferred compensation benefits to our employees, including our named executive officers.
All of our full-time employees, including our named executive officers, are eligible to participate in our health and welfare plans, including medical, dental, and vision benefits, short-term and long-term disability insurance, basic life and accidental death and dismemberment insurance, and certain voluntary benefits.
Employment Arrangements for Named Executive Officers
We initially entered into employment agreements or offer letters with each of our named executive officers in connection with their commencement of employment, which set forth the terms and conditions of their employment, including base salary, target annual bonus opportunity, initial equity awards, severance benefits and eligibility to participate in our employee benefit plans generally offered to our employees. The material terms of the offer letters are summarized below. In connection with this offering, we have adopted a new executive severance plan (the “Executive Severance Plan”), which will become effective upon the closing of this offering. Each of the NEOs may participate in the Executive Severance Plan, and it will supersede the severance provisions in the NEOs’ offer letters, if any. A detailed description of the Executive Severance Plan is provided in the section titled “Employee Benefit and Equity Compensation Plans.”
Current Offer Letters
Andrew (Andy) Orth, M.B.A.
On November 9, 2024, we entered into an offer letter with Mr. Orth for the position of President and Chief Executive Officer (the “Orth Offer Letter”), which provides for Mr. Orth’s at-will employment. Pursuant to the Orth Offer Letter, Mr. Orth is eligible to receive an annual base salary, subject to adjustment pursuant to our employee compensation policies in effect from time to time, and an annual performance bonus, as determined by our board of directors based on objective or subjective criteria approved by our board of directors, each of which has subsequently been increased as described above. Mr. Orth is also eligible to participate in the employee benefit plans generally available to our employees, subject to the terms of those plans.
Pursuant to the Orth Offer Letter, Mr. Orth received four option grants to purchase an aggregate of 1,504,670 shares of our common stock. The first option grant covers 944,538 shares and is subject to time-based vesting, with 25% of the shares subject to the option vesting after 12 months of continuous service from Mr. Orth’s start date and the remaining shares vesting in equal monthly installments over the following 36 months. The second option grant covers 404,174 shares and vests upon a specified regulatory development milestone, subject to Mr. Orth’s continuous service through such approval. The third option grant covers 77,979 shares and vests upon the closing of our initial public offering, subject to certain financial conditions as set forth in the Orth Offer Letter and Mr. Orth’s continuous service through such closing. The fourth option grant covers 77,979 shares and vests upon the successful completion of a clinical trial that satisfies the criteria set forth in the Orth Offer Letter, subject to Mr. Orth’s continuous service through such completion. In addition, Mr. Orth will immediately vest in his remaining unvested options if (i) the Company is subject to a “change in control” before Mr. Orth’s service with the Company terminates and (ii) upon or within 12 months after such change in control, Mr. Orth’s employment is terminated by the Company without “cause” or due to Mr. Orth’s “resignation for good reason,” in each case, as defined in the Orth Offer Letter.
In addition, pursuant to the Orth Offer Letter, in the event that Mr. Orth’s employment is terminated by us without “cause” or due to Mr. Orth’s “resignation for good reason,” subject to Mr. Orth’s execution of a general release of claims in favor of us and satisfaction of certain other conditions, Mr. Orth is entitled to receive (i) continuation of his then-current base salary for a period of nine months and (ii) up to nine months of Consolidated Omnibus Budget Reconciliation Act (“COBRA”) premium payments at the same rate we pay for active employees and their eligible dependents.
Baisong Mei, M.D., Ph.D.
On March 21, 2025, we entered into an offer letter with Dr. Mei for the position of Chief Medical Officer (the “Mei Offer Letter”), which provides for Dr. Mei’s at-will employment. Pursuant to the Mei Offer Letter, Dr. Mei is eligible to receive an annual base salary, subject to adjustment pursuant to our employee compensation policies in effect from time to time, and an annual performance bonus, as determined by our Chief Executive Officer and approved by our board of directors based on objective or subjective criteria, each of which has subsequently been increased as described above. Dr. Mei is also eligible to participate in the employee benefit plans generally available to our employees, subject to the terms of those plans. Dr. Mei also received a sign-on bonus of $50,000, subject to repayment if his employment had terminated within one year of his start date for any reason other than a termination by us without “cause” or by Dr. Mei’s “resignation for good reason,” in each case as defined in the Mei Offer Letter.
Pursuant to the Mei Offer Letter, Dr. Mei received two option grants to purchase an aggregate of 360,681 shares of our common stock. The first option grant covers 252,477 shares and is subject to time-based vesting, with 25% of the shares subject to the option vesting after 12 months of continuous service from Dr. Mei’s start date and the remaining shares vesting in equal monthly installments over the following 36 months. The second option grant covers 108,204 shares and vests upon achievement of certain specified clinical development milestone based on clinical proof-of-concept data, in each case as determined by our board of directors, subject to Dr. Mei’s continuous service through such date. In addition, Dr. Mei will immediately vest in his remaining unvested option if (i) the Company is subject to a “change in control” before Dr. Mei’s service with the Company terminates and (ii) upon or within 12 months after such change in control, Dr. Mei’s employment is terminated by the Company without “cause” or due to Dr. Mei’s “resignation for good reason,” in each case, as defined in the Mei Offer Letter.
In addition, pursuant to the Mei Offer Letter, in the event that Dr. Mei’s employment is terminated by us without “cause” or due to Dr. Mei’s “resignation for good reason,” subject to Dr. Mei’s execution of a general release of claims in favor of us and satisfaction of certain other conditions, Dr. Mei is entitled to receive (i) continuation of his then-current base salary for a period of nine months and (ii) up to nine months of COBRA premium payments at the same rate we pay for active employees and their eligible dependents.
Tracy Zimmermann, Ph.D.
On May 8, 2024, we entered into an offer letter with Dr. Zimmermann for the position of Chief Scientific Officer (the “Zimmermann Offer Letter”), which provides for Dr. Zimmermann’s at-will employment. Pursuant to the Zimmermann Offer Letter, Dr. Zimmermann is eligible to receive an annual base salary, subject to adjustment pursuant to our employee compensation policies in effect from time to time, and an annual performance bonus, as determined by our Chief Executive Officer and approved by our board of directors based on objective or subjective criteria, each of which has subsequently been increased as described above. Dr. Zimmermann is also eligible to participate in the employee benefit plans generally available to our employees, subject to the terms of those plans.
Pursuant to the Zimmermann Offer Letter, Dr. Zimmermann received three option grants to purchase an aggregate of 449,083 shares of our common stock. The first option grant covers 251,487 shares and is subject to time-based vesting, with 25% of the shares subject to the option vesting after 12 months of continuous service from Dr. Zimmermann’s start date and the remaining shares vesting in equal monthly installments over the following 36 months. The second option grant covers 107,780 shares and vests upon a specified regulatory development milestone, subject to Dr. Zimmermann’s continuous service through such approval. The third option grant covers 89,816 shares and vests upon achievement
of a specified clinical development milestone based on clinical proof-of-concept data. In addition, Dr. Zimmermann will immediately vest in her remaining unvested option if (i) the Company is subject to a “change in control” before Dr. Zimmermann’s service with the Company terminates and (ii) upon or within 12 months after such change in control, Dr. Zimmermann’s employment is terminated by the Company without “cause” or due to Dr. Zimmermann’s “resignation for good reason,” in each case, as defined in the Zimmermann Offer Letter.
In addition, pursuant to the Zimmermann Offer Letter, in the event that Dr. Zimmermann’s employment is terminated by us without “cause” or due to Dr. Zimmermann’s “resignation for good reason,” subject to Dr. Zimmermann’s execution of a general release of claims in favor of us and satisfaction of certain other conditions, Dr. Zimmermann is entitled to receive (i) continuation of her then-current base salary for a period of nine months and (ii) up to nine months of COBRA premium payments at the same rate we pay for active employees and their eligible dependents.
Outstanding Equity Awards at 2025 Fiscal Year-End
The following table sets forth information concerning outstanding equity awards held by each of our named executive officers as of December 31, 2025.
| |
|
|
|
|
|
Option Awards(1)
|
|
|
Name
|
|
|
Grant Date
|
|
|
Number of
Securities
Underlying
Unexercised
Options (#)
Exercisable
|
|
|
Number of
Securities
Underlying
Unexercised
Options (#)
Unexercisable
|
|
|
Equity Incentive
Plan Awards
Number of
Securities
Underlying
Unexercised
Unearned
Options
(#)
|
|
|
Option
Exercise
Price
($)
|
|
|
Option
Expiration
Date
|
|
|
Andrew Orth, M.B.A.
|
|
|
12/11/2024(2)
|
|
|
|
|
236,134 |
|
|
|
|
|
708,404 |
|
|
|
|
|
— |
|
|
|
|
|
0.92 |
|
|
|
|
|
12/10/2034 |
|
|
| |
12/11/2024(3)
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
|
|
404,174 |
|
|
|
|
|
0.92 |
|
|
|
|
|
12/10/2034 |
|
|
| |
12/11/2024(4)
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
|
|
77,979 |
|
|
|
|
|
0.92 |
|
|
|
|
|
12/10/2034 |
|
|
| |
12/11/2024(5)
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
|
|
77,979 |
|
|
|
|
|
0.92 |
|
|
|
|
|
12/10/2034 |
|
|
|
Baisong Mei, M.D., Ph.D.
|
|
|
05/13/2025(2)
|
|
|
|
|
— |
|
|
|
|
|
252,477 |
|
|
|
|
|
— |
|
|
|
|
|
1.84 |
|
|
|
|
|
05/12/2034 |
|
|
| |
05/13/2025(6)
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
|
|
108,204 |
|
|
|
|
|
1.84 |
|
|
|
|
|
05/12/2034 |
|
|
|
Tracy Zimmermann, Ph.D.
|
|
|
07/16/2024(2)
|
|
|
|
|
94,307 |
|
|
|
|
|
157,179 |
|
|
|
|
|
— |
|
|
|
|
|
0.92 |
|
|
|
|
|
07/15/2034 |
|
|
| |
07/16/2024(7)
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
|
|
107,780 |
|
|
|
|
|
0.92 |
|
|
|
|
|
07/15/2034 |
|
|
| |
07/16/2024(8)
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
|
|
89,816 |
|
|
|
|
|
0.92 |
|
|
|
|
|
07/15/2034 |
|
|
| |
02/25/2025(9)
|
|
|
|
|
1,200 |
|
|
|
|
|
4,565 |
|
|
|
|
|
— |
|
|
|
|
|
0.92 |
|
|
|
|
|
02/24/2035 |
|
|
| |
02/25/2025(10)
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
|
|
13,454 |
|
|
|
|
|
0.92 |
|
|
|
|
|
02/24/2035 |
|
|
(1)
All stock option awards were granted under our 2023 Plan, described below. All options are exercisable immediately upon grant, subject to a repurchase right in favor of the Company, which lapses as the option vests. Accordingly, the columns and footnotes below reflect the extent to which the stock options held by our named executive officers were vested (as opposed to exercisable) as of December 31, 2025.
(2)
The shares underlying this stock option award vest as follows: 25% of the shares vested on grant date, and the remaining 75% of the shares vest in 36 equal monthly installments thereafter, subject to the executive’s continued service relationship through the applicable vesting date.
(3)
The shares underlying this stock option award vest as follows: immediately upon the Company’s first approval of a specified regulatory development milestone, subject to Mr. Orth’s continued service relationship through the applicable vesting date.
(4)
The shares underlying this stock option award vest as follows: immediately upon the Company’s closing of an initial public offering, subject to certain financial conditions as set forth in the Orth Offer Letter and Mr. Orth’s continuous service through such closing date.
(5)
The shares underlying this stock option award vest as follows: immediately upon the Company’s successful completion of specified clinical development milestones, subject to Mr. Orth’s continuous service through the applicable vesting date.
(6)
The shares underlying this stock option award vest as follows: immediately upon achievement of certain specified clinical development milestone based on clinical proof-of-concept data, subject to Dr. Mei’s continuous service through the applicable vesting date.
(7)
The shares underlying this stock option award vest as follows: immediately upon a specified regulatory development milestone, subject to Dr. Zimmermann’s continuous service through the applicable vesting date.
(8)
The shares underlying this stock option award vest as follows: immediately upon the achievement of a specified clinical development milestone, subject to Dr. Zimmermann’s continuous service through the applicable vesting date.
(9)
The shares underlying this stock option award vest in 48 equal monthly installments thereafter, subject to Dr. Zimmermann’s continued service relationship through the applicable vesting date.
(10)
The shares underlying this stock option award vest as follows: immediately upon achievement of a specified clinical development milestone, subject to Dr. Zimmermann’s continuous service through the applicable vesting date.
Employee Benefit and Equity Compensation Plans
2023 Stock Plan
The 2023 Plan was initially adopted by our board of directors on July 17, 2023, and approved by our stockholders on August 14, 2023. The 2023 Plan was subsequently amended on April 16, 2024, December 11, 2024, February 25, 2025, January 23, 2026, May 20, 2026, and June 3, 2026 to increase the number of shares reserved for issuance thereunder. Under the 2023 Plan, as amended, we have reserved for issuance an aggregate of 10,017,776 shares of our common stock. These numbers are subject to adjustment in the event of certain changes in our capitalization, including a subdivision of our outstanding common stock, a dividend payable in shares of common stock, a combination or consolidation of our outstanding common stock into a lesser number of shares, a reclassification, or other increase or decrease in the number of issued shares of our common stock effected without receipt of consideration by us. Our board of directors has determined not to issue any further awards under the 2023 Plan following the completion of this offering, but all outstanding awards under the 2023 Plan will continue to be governed by their existing terms. In connection with this offering, we intend to adopt a new equity incentive plan under which we will grant equity-based awards following this offering, as described below under “2026 Stock Option and Incentive Plan.” The following summary describes the material terms of the 2023 Plan. This summary is not a complete description of all provisions of the 2023 Plan and is qualified in its entirety by reference to the 2023 Plan, which will be filed as an exhibit to the registration statement of which this prospectus is a part.
The shares of common stock previously issued under the 2023 Plan that are forfeited to or repurchased by us due to failure to vest are currently added back to the shares of common stock available for issuance under the 2023 Plan. In addition, shares that otherwise would have been issuable under the 2023 Plan but are withheld by us in payment of the purchase price or exercise price of an award or in satisfaction of withholding taxes remain available for issuance under the 2023 Plan. Shares subject to any outstanding option, restricted stock unit or other right that expires or is cancelled for any reason, to the extent allocable to the unexercised or unsettled portion of such award, remain available for issuance under the 2023 Plan. To the extent an award is settled in cash, the cash settlement does not reduce the number of shares remaining available for issuance under the 2023 Plan. Following the completion of this offering, any such shares that would otherwise become available for issuance under the 2023 Plan will instead be added to the shares of common stock available for issuance under the 2026 Plan (as defined below).
Our board of directors has acted as administrator of the 2023 Plan. The administrator has full authority and discretion to take any actions it deems necessary or advisable for the administration of the 2023 Plan. Persons eligible to participate in the 2023 Plan are our employees, outside directors and consultants.
The 2023 Plan permits the granting of (i) options to purchase common stock intended to qualify as incentive stock options under Section 422 of the Code and (ii) options that do not so qualify. The option exercise price of each option is determined by the administrator but may not be less than 100% of the fair market value of our common stock on the date of grant or, in the case of an incentive stock option granted to a 10% owner, the exercise price shall not be less than 110% of the fair market value of our common stock on the date of grant. The term of each option is fixed by the administrator and may not
exceed ten years from the date of grant, or five years in the case of certain incentive stock options grants. The administrator determines when each option becomes vested and exercisable.
The administrator of the 2023 Plan may award or sell shares of common stock subject to such terms and conditions as it determines, including forfeiture conditions, repurchase rights, rights of first refusal and other transfer restrictions.
The administrator of the 2023 Plan may also grant restricted stock units subject to such vesting conditions and other terms and conditions as it determines. Restricted stock units may be settled in shares of common stock, cash or a combination of both, as determined by the administrator.
In the event of certain changes in our capitalization, including a subdivision of our outstanding common stock, a dividend payable in shares of common stock, a combination or consolidation of our outstanding common stock into a lesser number of shares, a reclassification or any other increase or decrease in the number of issued shares of common stock effected without receipt of consideration by us, proportionate adjustments will be made, as applicable, to the number and kind of shares available for issuance under the 2023 Plan, the number and kind of shares subject to outstanding awards and outstanding rights to purchase shares, the exercise price or purchase price applicable to such awards or rights and any applicable repurchase price. In addition, in the event of an extraordinary dividend payable in a form other than shares, a recapitalization, a spin-off or similar occurrence, the administrator may make appropriate adjustments to such items.
The 2023 Plan provides that, in the event that the Company is a party to a merger or consolidation, or in the event of a sale of all or substantially all of our stock or assets, shares acquired under the 2023 Plan and outstanding awards under the 2023 Plan will be treated in the manner described in the definitive transaction agreement or, if there is no such agreement to which the Company is a party, as determined by our board of directors. Such treatment need not be the same for all awards or all portions of an award and may include, among other things: (i) continuing or assuming the award or substituting a comparable award, (ii) cancelling the award and making a payment with respect to the vested portion of such award, (iii) cancelling an option without payment following notice and an opportunity to exercise the option to the extent vested or becoming vested in connection with the transaction, (iv) suspending option exercise rights for a limited period prior to the closing of the transaction and/or (v) terminating any early exercise rights. Our board of directors also has discretion to accelerate, in whole or in part, the vesting and exercisability of an award in connection with such corporate transaction.
Our board of directors may amend, suspend or terminate the 2023 Plan at any time, subject to stockholder approval where required by applicable law. Within the limitations of the 2023 Plan, our board of directors may modify, reprice, extend or assume outstanding options or may accept the cancellation of outstanding options in return for the grant of new options or a different type of award for the same or a different number of shares and at the same or a different exercise price, if applicable. No modification of an option may, without the consent of the optionee, impair the optionee’s rights or increase the optionee’s obligations under such option; provided, however, that a modification of an option that is otherwise favorable to the optionee will not require the consent of the optionee. The Company may also cancel certain options that were not granted in compliance with Rule 701 under the Securities Act upon not less than 30 days’ prior written notice to the optionee. If the Company elects to cancel such option, it will deliver to the optionee consideration with an aggregate value equal to the excess of the fair market value of the shares subject to such option as of the time of the cancellation over the exercise price of such option. If the consideration would be a negative amount, such option may be cancelled without the delivery of any consideration.
No awards may be granted under our 2023 Plan after the date that is ten years from the later of (i) the date our 2023 Plan was adopted by our board of directors or (ii) the date our board of directors approved the most recent increase in the number of shares reserved under our 2023 Plan that was also approved by our stockholders. As described above, our board of directors has determined not to issue any further awards under our 2023 Plan following the completion of this offering.
2026 Stock Option and Incentive Plan
The 2026 Stock Option and Incentive Plan (the “2026 Plan”) was adopted by our board of directors on September 19, 2026, was approved by our stockholders on October 8, 2026, and will become effective
on the date immediately preceding the date on which the registration statement of which this prospectus is a part is declared effective by the SEC. The 2026 Plan will allow us to make equity-based and cash-based incentive awards to our officers, employees, directors, and consultants. The following summary describes the material terms of the 2026 Plan. This summary is not a complete description of all provisions of the 2026 Plan and is qualified in its entirety by reference to the 2026 Plan, which will be filed as an exhibit to the registration statement of which this prospectus is a part.
Authorized Shares. We have initially reserved 5,033,227 shares of our common stock for the issuance of awards under the 2026 Plan (the “Initial Limit”), plus the number of shares reserved and remaining available for issuance under the 2023 Plan as of the effective date of the 2026 Plan. The number of shares reserved and available for issuance under the 2026 Plan will automatically increase on January 1, 2027 and each January thereafter during the term of the 2026 Plan, by 5% of the outstanding number of shares of our common stock on the immediately preceding December 31 (including, for this purpose, shares underlying any outstanding prefunded warrants) or such lesser number of shares as determined by our culture and compensation committee (the “Annual Increase”). The number of shares reserved for issuance under the 2026 Plan will be subject to adjustment in the event of a stock split, stock dividend, or other change in our capitalization. The shares we issue under the 2026 Plan will be authorized but unissued shares or shares that we reacquire.
The shares of common stock underlying any awards under the 2026 Plan or the 2023 Plan that are forfeited, cancelled, held back upon exercise or settlement of an award to satisfy the exercise price or tax withholding, reacquired by us prior to vesting, satisfied without the issuance of stock, or are otherwise terminated (other than by exercise) will be added back to the shares of common stock available for issuance under the 2026 Plan. The maximum number of shares of common stock that may be issued pursuant to incentive stock options shall not exceed the Initial Limit, cumulatively increased on January 1, 2027 and on each January 1 thereafter by the lesser of the Annual Increase for such year or 5,033,227 shares of common stock.
Non-Employee Director Compensation Limit. The grant date fair value of all awards under the 2026 Plan and all other cash compensation paid by us to any non-employee director during any one calendar year for services as a non-employee director may not exceed $750,000; provided, however, that such amount shall be $1,000,000 for the calendar year in which the applicable non-employee director is initially elected or appointed to our board of directors.
Plan Administration. The 2026 Plan will be administered by our culture and compensation committee. Our culture and compensation committee will have full power to select, from among the individuals eligible for awards, the individuals to whom awards will be granted and the number of shares subject to such awards, to make any combination of awards to participants, to accelerate at any time the exercisability or vesting of any award, and to determine the specific terms and conditions of each award, subject to the provisions of the 2026 Plan. The culture and compensation committee is specifically authorized to exercise its discretion to reduce the exercise price of outstanding stock options and stock appreciation rights or effect the repricing of such awards through cancellation and re-grants without stockholder consent.
Eligibility. Persons eligible to participate in the 2026 Plan will be those employees, non-employee directors, and consultants selected from time to time by our culture and compensation committee in its discretion.
Stock Options. The 2026 Plan permits the granting of both options to purchase common stock intended to qualify as incentive stock options under Section 422 of the Code and options that do not so qualify. The option exercise price of each option will be determined by our culture and compensation committee but may not be less than 100% of the fair market value of our common stock on the date of grant unless the option (i) is granted pursuant to a transaction described in, and in a manner consistent with, Section 424(a) of the Code, (ii) is granted to an individual who is not subject to United States income tax, or (iii) complies with or is exempt from Section 409A of the Code. The term of each option will be fixed by our culture and compensation committee and may not exceed ten years from the date of grant (or five years in the case of certain incentive stock options). Our culture and compensation committee will determine at what time or times each option may be exercised.
Stock Appreciation Rights. Our culture and compensation committee may award stock appreciation rights under the 2026 Plan subject to such conditions and restrictions as it determines. Stock appreciation rights entitle the recipient to shares of common stock, or cash, equal to the value of the appreciation in our stock price over the exercise price. The exercise price of each stock appreciation right may not be less than 100% of the fair market value of our common stock on the date of grant unless the stock appreciation right (i) is granted pursuant to a transaction described in, and in a manner consistent with, Section 424(a) of the Code, (ii) is granted to an individual who is not subject to United States income tax, or (iii) complies with or is exempt from Section 409A of the Code. The term of each stock appreciation right will be fixed by our culture and compensation committee and may not exceed ten years from the date of grant. Our culture and compensation committee will determine at what time or times each stock appreciation right may be exercised.
Restricted Stock and Restricted Stock Units. Our culture and compensation committee may award restricted shares of common stock and restricted stock units to participants subject to such conditions and restrictions as it determines. These conditions and restrictions may include the achievement of certain performance goals and/or continued employment or other service relationship with us through a specified vesting period.
Unrestricted Stock Awards. Our culture and compensation committee may grant shares of common stock that are free from any restrictions under the 2026 Plan. Unrestricted stock may be granted to participants in recognition of past services or for other valid consideration and may be issued in lieu of cash compensation due to such participant.
Dividend Equivalent Rights. Our culture and compensation committee may grant dividend equivalent rights to participants that entitle the recipient to receive credits for dividends that would be paid if the recipient had held a specified number of shares of common stock.
Cash-Based Awards. Our culture and compensation committee may grant cash bonuses under the 2026 Plan to participants, subject to the achievement of certain performance goals.
Sale Event. The 2026 Plan provides that, in the case of and subject to the consummation of a “sale event” (as defined in the 2026 Plan), our culture and compensation committee may, on such terms and conditions as it deems appropriate and without a participant’s consent, take one or more of the following actions: (i) provide for the assumption or continuation of outstanding awards by the successor entity or the substitution of such awards with new awards of the successor entity or its parent, with appropriate adjustments; (ii) provide that an award will vest and become exercisable, realizable or payable, or that restrictions applicable to an award will lapse, in whole or in part, prior to or upon consummation of the sale event and, to the extent determined by our culture and compensation committee, terminate upon or immediately prior to the effectiveness of the sale event; (iii) permit individuals holding options and stock appreciation rights to exercise such options and stock appreciation rights, to the extent then vested and exercisable, within a specified period prior to the consummation of the sale event, following which any unexercised awards will terminate; (iv) make or provide for a payment, in cash or in kind, to participants holding vested and exercisable options and stock appreciation rights, in exchange for the cancellation thereof, in an amount equal to the difference between the per share consideration payable to stockholders in the sale event and the applicable exercise price; provided that any options or stock appreciation rights with exercise prices equal to or greater than such per share consideration will be cancelled for no consideration; (v) make or provide for a payment, in cash or in kind, to participants holding other awards, in exchange for the cancellation thereof, in an amount equal to the per share consideration payable to stockholders in the sale event multiplied by the number of vested shares subject to such awards; (vi) terminate any award, or portion thereof, that is unvested immediately prior to the effective time of the sale event, with such payment to the participant, including no payment, as the our culture and compensation committee determines in its discretion; or (vii) take any combination of the foregoing actions. In taking any such actions, the plan administrator will not be required to treat all awards, all awards held by a participant or all awards of the same type in the same manner.
Amendment. Our board of directors may amend or discontinue the 2026 Plan and our culture and compensation committee may amend or cancel outstanding awards for purposes of satisfying
changes in law or any other lawful purpose, but no such action may materially and adversely affect rights under an award without the holder’s consent. Certain amendments to the 2026 Plan require the approval of our stockholders. The culture and compensation committee is specifically authorized to exercise its discretion to reduce the exercise price of outstanding stock options and stock appreciation rights or effect the repricing of such awards through cancellation and re-grants without stockholder consent.
No awards may be granted under the 2026 Plan after the date that is ten years from the effective date of the 2026 Plan. No awards have been made under the 2026 Plan prior to the date hereof.
2026 Employee Stock Purchase Plan
The 2026 Employee Stock Purchase Plan (the “ESPP”) was adopted by our board of directors on September 19, 2026, was approved by our stockholders on October 8, 2026, and will become effective on the date immediately preceding the date on which the registration statement of which this prospectus is part is declared effective by the SEC. The ESPP has two components: a component intended to qualify as an “employee stock purchase plan” within the meaning of Section 423 of the Code (the “423 Component”), and a component that is not intended to so qualify (the “Non-423 Component”). Except as otherwise provided, the Non-423 Component will be operated and administered in the same manner as the 423 Component, except where prohibited by law. The following summary describes the material terms of the ESPP. This summary is not a complete description of all provisions of the ESPP and is qualified in its entirety by reference to the ESPP, which will be filed as an exhibit to the registration statement of which this prospectus is a part.
Authorized Shares. The ESPP initially reserves and authorizes the issuance of up to a total of 503,322 shares of our common stock to participating employees. The ESPP provides that the number of shares reserved and available for issuance will automatically increase on January 1, 2027 and each January 1 thereafter through January 1, 2036, by the least of (i) 1% of the outstanding number of shares of our common stock on the immediately preceding December 31 (including, for this purpose, shares underlying any outstanding prefunded warrants), (ii) 1,509,966 shares of common stock, or (iii) such number of shares of common stock as determined by the administrator of the ESPP. The number of shares reserved under the ESPP is subject to adjustment in the event of a stock split, stock dividend, or other change in our capitalization.
Eligibility. All individuals classified as employees on the payroll records of the Company or a “designated company” (as defined in the ESPP) are generally eligible to participate in the ESPP, provided that the ESPP administrator may determine in advance of an offering that employees are eligible only if, as of the first day of the offering, they (a) have been employed by us or a designated company for such continuous period as the ESPP administrator may require, provided such service requirement is less than two years, and (b) are customarily employed by us or a designated company for more than 20 hours per week and more than five months per calendar year, or satisfy such other criteria as the ESPP administrator may determine consistent with Section 423 of the Code. However, an employee may not be granted an option under the ESPP if, immediately after the grant, the employee would be treated as owning stock possessing 5% or more of the total combined voting power or value of all classes of stock of the Company or any parent or subsidiary of the Company.
Offerings. We may make one or more offerings each year to our employees to purchase shares of our common stock under the ESPP, each of which may consist of one or more purchase periods. Offerings will begin and end on the dates determined by the ESPP administrator, except that no offering will exceed 27 months in duration. Each eligible employee may elect to participate in any offering by submitting an enrollment form by the deadline established by the ESPP administrator.
Each employee who is a participant in the ESPP may purchase shares by authorizing payroll deductions at a minimum of 1% and up to a maximum of 15% of such participant’s eligible compensation during an offering period (or such other minimum and maximum as determined by the administrator in advance of an offering). Unless the participating employee has previously withdrawn from the offering, such participant’s accumulated payroll deductions or contributions will be used to purchase shares of our common stock on the last day of each purchase period at a price equal to 85% of the fair market value
of the shares on the first day or the last day of the purchase period, whichever is lower, provided that no more than the number of shares of common stock determined by dividing $25,000 by the fair market value of our common stock on the first day of such offering period (or such other maximum number of shares as may be established by the administrator) may be purchased by any one employee during any purchase period. Under applicable tax rules, an employee may purchase no more than $25,000 worth of shares of our common stock, valued at the start of the offering period, under the ESPP for each calendar year during which any option granted to the employee is outstanding at any time.
The accumulated payroll deductions of any employee who is not a participant on the last day of an offering period will be refunded. An employee’s rights under the ESPP terminate upon voluntary withdrawal from the plan or when the employee ceases employment with us for any reason.
Sale Event. In the case of and subject to the consummation of a “sale event” (as defined in the ESPP), the administrator of the ESPP, in its discretion, and on such terms and conditions as it deems appropriate, is authorized to take any one or more of the following actions under the ESPP or with respect to any right under the ESPP to facilitate such transactions or events: (i) provide for either (A) termination of any outstanding option in exchange for an amount of cash, if any, equal to the amount that would have been obtained upon the exercise of such option had such option been currently exercisable or (B) the replacement of such outstanding option with other options or property selected by the administrator of the ESPP in its sole discretion; (ii) provide that the outstanding options under the ESPP shall be assumed by the successor or survivor corporation, or a parent or subsidiary thereof, or shall be substituted for similar options covering the stock of the successor or survivor corporation, or a parent or subsidiary thereof, with appropriate adjustments as to the number and kind of shares and prices; (iii) make adjustments in the number and type of shares of common stock (or other securities or property) subject to outstanding options under the ESPP and/or in the terms and conditions of outstanding options and options that may be granted in the future; (iv) provide that the offering with respect to which an option relates will be shortened by setting a new exercise date on which such offering period will end; and (v) provide that all outstanding options shall terminate without being exercised and all amounts in the accounts of participants shall be promptly refunded.
Amendment. The ESPP may be terminated or amended by our board of directors at any time. An amendment that increases the number of shares of our common stock authorized under the ESPP and certain other amendments require the approval of our stockholders.
Senior Executive Cash Incentive Bonus Plan
On September 19, 2026, our board of directors adopted the Senior Executive Cash Incentive Bonus Plan (the “Bonus Plan”), which will become effective upon the effectiveness of the registration statement of which this prospectus forms a part. The Bonus Plan provides for cash bonus payments based upon company and individual performance targets established by our culture and compensation committee. The payment targets will be related to financial and operational measures or objectives with respect to our company, or the corporate performance goals, as well as individual performance objectives. The following summary describes the material terms of the Bonus Plan. This summary is not a complete description of all provisions of the Bonus Plan and is qualified in its entirety by reference to the Bonus Plan, which will be filed as an exhibit to the registration statement of which this prospectus is a part.
Our culture and compensation committee may select corporate performance goals from among the following: research and development, publication, clinical and/or regulatory milestones; cash flow (including, but not limited to, operating cash flow and free cash flow); revenue; earnings before interest, taxes, depreciation and amortization; net income (loss) (either before or after interest, taxes, depreciation and/or amortization); changes in the market price of our common stock; economic value-added; acquisitions, licenses, collaborations or strategic transactions, including government or other third-party grants, asset sales, royalty financings, partnership, collaboration and licensing arrangements, or a combination of these approaches; financing or other capital raising transactions; operating income (loss); return on capital, assets, equity, or investment; stockholder returns; return on sales; gross or net profit levels; productivity; expense efficiency; margins; operating efficiency; working capital; earnings (loss) per share of our common stock; sales or market shares; number of prescriptions
or prescribing physicians; coverage decisions; leadership development, employee retention and recruiting, and other human resources matters; operating income and/or net annual recurring revenue, or any other performance goal selected by the culture and compensation committee, any of which may be (A) measured in absolute terms, as compared to any incremental increase, (B) measured in terms of growth, (C) compared to another company or companies or to results of a peer group, (D) measured against the market as a whole and/or as compared to applicable market indices and/or (E) measured on a pre-tax or post-tax basis, as applicable. Corporate performance goals may be used to measure the performance of our company as a whole, a business unit or other segment, or one or more product lines or specific markets.
Each executive officer who is selected to participate in the Bonus Plan will have a target bonus opportunity set for each performance period. The bonus formulas will be adopted in each performance period by the culture and compensation committee and communicated to each executive at the beginning of the performance period. The corporate performance goals will be measured at the end of the fiscal year after our financial reports have been published. Subject to the rights contained in any agreement between the executive officer and us, an executive officer shall be required to be employed by us on the bonus payment date to be eligible to receive a bonus payment under the Bonus Plan. If an executive officer was not employed for the entire performance period, the culture and compensation committee may prorate the bonus based on the number of days employed during the period. Bonus payments will be made as soon as practicable, but not later than two and one-half months after the end of the fiscal year in which the applicable performance period ends. The Bonus Plan also permits the culture and compensation committee to approve additional bonuses to executive officers in its sole discretion.
Executive Severance Plan
On September 19, 2026, our board of directors adopted the City Therapeutics, Inc. Executive Severance Plan (the “Severance Plan”), which will become effective upon the effectiveness of the registration statement of which this prospectus forms a part. Our Chief Executive Officer will participate in the Severance Plan as a Tier 1 executive; our Chief Financial Officer, Chief Medical Officer, Chief Scientific Officer, Chief Operating Officer, Chief Legal Officer and Chief Human Resources Officer will participate as Tier 2 executives; and our employees at the senior vice president level will participate as Tier 3 executives, in each case subject to the execution of a participation agreement. The benefits provided under the Severance Plan will replace any other severance or change-in-control payments and benefits for which a participating executive may otherwise be eligible under an existing offer letter, employment agreement or other arrangement.
Under the Severance Plan, if a participating executive’s employment is terminated by us other than for cause, death or disability, or the executive resigns for good reason, in either case outside the period beginning three months before and ending 12 months after a change in control, the executive will be eligible to receive (i) an amount equal to 12 months of base salary for a Tier 1 or Tier 2 executive and nine months of base salary for a Tier 3 executive, payable in substantially equal installments, and (ii) subject to the executive’s timely election of continued health coverage under COBRA, payment of the employer portion of the executive’s COBRA premiums for up to 12 months for a Tier 1 or Tier 2 executive and nine months for a Tier 3 executive.
If such a termination occurs during the period beginning three months before and ending 12 months after a change in control, the executive will instead be eligible to receive (i) a lump-sum payment equal to 150% of base salary and target annual bonus for a Tier 1 executive, 100% of base salary and target annual bonus for a Tier 2 executive and 75% of base salary and target annual bonus for a Tier 3 executive; (ii) subject to the executive’s timely election of continued health coverage under COBRA, payment of the executive’s COBRA premiums for up to 18 months for a Tier 1 executive, 12 months for a Tier 2 executive and nine months for a Tier 3 executive; and (iii) full accelerated vesting of all outstanding equity awards subject solely to time-based vesting.
The foregoing severance payments and benefits are subject to the participating executive’s timely execution and non-revocation of a separation agreement and release of claims in our favor. If any payments or benefits payable to a participating executive would be subject to the excise tax imposed by Section 4999 of the Code, such payments and benefits will be reduced to the extent necessary to avoid the excise tax if such reduction would result in the executive receiving a greater after-tax benefit.
DIRECTOR COMPENSATION
2025 Director Compensation Table
The following table presents the compensation awarded to, earned by, or paid to each person who served as a non-employee member of our board of directors for their services to us during Fiscal Year 2025. Other than as set forth in the table and described more fully below, we did not pay any compensation, make any equity awards or non-equity awards to, or pay any other compensation to any of the members of our board of directors in Fiscal Year 2025 for their services as members of the board of directors. During Fiscal Year 2025, Mr. Orth served as a member of our board of directors but received no additional compensation for his service as a member of our board of directors. The compensation for Fiscal Year 2025 received by Mr. Orth as our President and Chief Executive Officer is presented in the section titled “Executive Compensation — 2025 Summary Compensation Table” above.
|
Name
|
|
|
Fees Earned or
Paid in Cash
($)(1)
|
|
|
Option Awards
($)(2)(3)
|
|
|
Total
($)
|
|
|
John Maraganore, Ph.D.
|
|
|
|
|
200,000 |
|
|
|
|
|
2,325,602 |
|
|
|
|
|
2,525,602 |
|
|
|
Barry Greene
|
|
|
|
|
50,000 |
|
|
|
|
|
205,026 |
|
|
|
|
|
255,026 |
|
|
|
Robert Nelsen, M.B.A.
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
Saraswathy (Sara) V. Nochur, Ph.D.(4)
|
|
|
|
|
49,059 |
|
|
|
|
|
205,935 |
|
|
|
|
|
254,994 |
|
|
|
Ron Philip
|
|
|
|
|
50,000 |
|
|
|
|
|
205,982 |
|
|
|
|
|
255,982 |
|
|
(1)
The amounts reported represent the cash fees each director received for their services to our board of directors during Fiscal Year 2025.
(2)
The amounts reported represent the aggregate grant date fair value of stock option awards granted to our directors in Fiscal Year 2025, computed in accordance with FASB ASC Topic 718. Such grant date fair values do not take into account any estimated forfeitures related to service-based vesting. The assumptions used in calculating the grant date fair values of the option awards reported in this column are set forth in Note 12 of our consolidated financial statements for Fiscal Year 2025, included elsewhere in this prospectus. The amounts reported in this column reflect the accounting cost for these option awards and do not correspond to the actual economic value that may be received by our directors upon the exercise of the option awards or any sale of the underlying securities.
(3)
As of December 31, 2025, each non-employee director held options to purchase the aggregate number of shares of our common stock as set forth below. As of December 31, 2025, none of the non-employee directors held any unvested stock awards, except that Dr. Maraganore held 815,530 shares of restricted common stock.
(4)
Dr. Nochur was appointed as a director on February 25, 2025. The fees earned by or paid to Dr. Nochur for Fiscal Year 2025 were prorated based on her date of appointment.
|
Name
|
|
|
Shares Underlying
Outstanding
Option Awards
at Fiscal Year-End
|
|
|
John Maraganore, Ph.D.
|
|
|
|
|
— |
|
|
|
Barry Greene
|
|
|
|
|
59,878 |
|
|
|
Robert Nelsen, M.B.A.
|
|
|
|
|
— |
|
|
|
Saraswathy (Sara) V. Nochur, Ph.D.
|
|
|
|
|
59,878 |
|
|
|
Ron Philip
|
|
|
|
|
59,878 |
|
|
Consulting Agreement with Dr. Maraganore
On August 14, 2023, we entered into a consulting agreement with Dr. Maraganore, pursuant to which Dr. Maraganore provides certain consulting and advisory services to us. For additional information regarding Dr. Maraganore’s consulting agreement, see “Certain Relationships and Related Person Transactions — Maraganore Consulting Agreement.”
Executive Chair Agreement with Dr. Maraganore
On November 10, 2023, we entered into an executive chair agreement with Dr. Maraganore, pursuant to which Dr. Maraganore agreed to provide services as Executive Chair and a member of our board of directors. For additional information regarding Dr. Maraganore’s executive chair agreement, see “Certain Relationships and Related Person Transactions — Maraganore Executive Chair Agreement.”
Non-Employee Director Compensation Policy
In connection with this offering, our board of directors has adopted a non-employee director compensation policy, which will become effective upon the effectiveness of the registration statement of which this prospectus forms a part. The policy will be designed to enable us to attract and retain, on a long-term basis, high-caliber non-employee directors who are not employees or officers of us or our subsidiaries. Unless otherwise determined by our board of directors, non-employee directors who are employees of, or otherwise affiliated with, an institutional investor in us will not be eligible to receive compensation under the policy. Under the policy, each director who is not an employee will be paid cash compensation from and after the completion of this offering, as set forth below:
|
Board of Directors:
|
|
|
Annual
Retainer ($)
|
|
|
Members
|
|
|
|
$ |
40,000 |
|
|
|
Additional retainer for non-executive chair
|
|
|
|
$ |
30,000 |
|
|
|
Audit Committee:
|
|
|
|
|
|
|
|
|
Members (other than chair)
|
|
|
|
$ |
10,000 |
|
|
|
Retainer for chair
|
|
|
|
$ |
20,000 |
|
|
|
Culture and Compensation Committee:
|
|
|
|
|
|
|
|
|
Members (other than chair)
|
|
|
|
$ |
7,500 |
|
|
|
Retainer for chair
|
|
|
|
$ |
15,000 |
|
|
|
Nominating and Corporate Governance Committee:
|
|
|
|
|
|
|
|
|
Members (other than chair)
|
|
|
|
$ |
5,000 |
|
|
|
Retainer for chair
|
|
|
|
$ |
10,000 |
|
|
In addition, the non-employee director compensation policy provides that, upon the effective date of the policy, each non-employee director will receive an option award to purchase 22,649 shares of our common stock (the “IPO Award”). The IPO Award will vest in full on the earlier of the first anniversary of the grant date or the next annual meeting of stockholders, subject to continued service through the applicable vesting date. Upon initial election or appointment to our board of directors, each non-employee director will be granted an initial option award to purchase 45,802 shares of our common stock (the “Initial Award”). The Initial Award will vest in equal monthly installments over three years from the grant date, subject to continued service through the applicable vesting date. Furthermore, on the date of each annual meeting of stockholders following the completion of this offering, each non-employee director who continues as a non-employee director following such meeting (other than a director receiving an Initial Award) will be granted an annual option award to purchase 22,649 shares of our common stock (the “Annual Award”). The Annual Award will vest on the earlier of the first anniversary of the grant date or the next annual meeting of stockholders, subject to continued service through the applicable vesting date. All outstanding Initial Awards and Annual Awards will become fully vested and exercisable upon a “sale event” (as defined in our 2026 Plan).
The aggregate amount of compensation, including both equity compensation and cash compensation, paid to any non-employee director for service as a non-employee director in a calendar year period will not exceed $750,000 (or $1,000,000 for the calendar year in which the applicable non-employee director is initially elected or appointed to our board of directors).
We will reimburse all reasonable out-of-pocket expenses incurred by non-employee directors in attending meetings of the board of directors and committees thereof. Employee directors will receive no additional compensation for their service as a director.
Equity Grants to Non-Employee Directors
In connection with this offering and the non-employee director compensation policy described above, our board of directors approved grants to each of our non-employee directors of options to purchase shares of common stock (the “Director IPO Grants”). The Director IPO Grants are contingent and subject to the effectiveness of the registration statement of which this prospectus forms a part. The Director IPO Grants will each have an exercise price per share equal to the per share “price to the public” (or equivalent) set forth on the cover page for the final prospectus relating to our initial public offering, expire ten years from the date of grant and vest in full on the earlier of the first anniversary of the grant date or the next annual meeting of stockholders, in each case subject to the applicable director’s continued service relationship through the vesting date. The Director IPO Grants will be subject to full accelerated vesting upon the sale of the Company.
CERTAIN RELATIONSHIPS AND RELATED PERSON TRANSACTIONS
In addition to the compensation arrangements, including employment, termination of employment and change in control arrangements, and indemnification arrangements discussed, when required, in the sections titled “Management,” “Director Compensation” and “Executive Compensation” and the registration rights described in the section titled “Description of Capital Stock — Registration Rights,” the following is a description of all transactions since January 1, 2024 and each currently proposed transaction in which:
•
we have been or are to be a participant;
•
the amounts involved exceeded or will exceed the lesser of $120,000 or 1% of the average of our total assets at the year-end for the last two completed fiscal years; and
•
any of our directors, executive officers or holders of more than 5% or more of our outstanding capital stock, or any immediate family member of, or person sharing the household with, any of these individuals or entities or affiliated entities, had or will have a direct or indirect material interest.
Convertible Notes
On January 5, 2024 and February 15, 2024, we issued two convertible notes with an aggregate principal amount of $0.5 million to John Maraganore, Ph.D., our co-founder, Executive Chair and a holder of more than 5% of our outstanding capital stock, in consideration for consulting services. The notes matured on the earlier of June 30, 2024 or the consummation of a deemed liquidation event. In connection with the closing of our Series A convertible preferred stock financing on April 17, 2024, the convertible notes converted into 56,095 shares of our Series A convertible preferred stock at a conversion price of $9.02655 per share, reflecting a 10% discount to the $10.0295 per share purchase price paid by the new investors in the Series A financing.
On January 5, 2024 and February 15, 2024, we issued two convertible notes with an aggregate principal amount of $4.5 million to ARCH Venture Fund XII, L.P., a holder of more than 5% of our outstanding capital stock. In connection with the closing of our Series A convertible preferred stock financing on April 17, 2024, the convertible notes converted into 504,869 shares of our Series A convertible preferred stock at a conversion price of $9.02655 per share, reflecting a 10% discount to the $10.0295 per share purchase price paid by the new investors in the Series A financing.
Series A Convertible Preferred Stock Financing
Between April 17, 2024 and December 30, 2024, we issued and sold an aggregate of 13,435,354 shares of our Series A convertible preferred stock at a purchase price of $10.0295 per share, for an aggregate purchase price of approximately $134.7 million, and issued an aggregate of 560,964 shares of Series A convertible preferred stock upon conversion of outstanding convertible notes.
The outstanding shares of Series A convertible preferred stock will convert into shares of our common stock at a rate of one share of common stock immediately prior to the completion of this offering. The following table summarizes the shares of our Series A convertible preferred stock issued to our related parties:
|
Purchasers
|
|
|
Shares of Series A
Convertible
Preferred Stock
|
|
|
Total
Purchase
Price
|
|
|
Total Convertible
Note Amount
|
|
|
ARCH Venture Fund XII, L.P.(1)
|
|
|
|
|
4,991,633 |
|
|
|
|
$ |
44,999,999.54 |
|
|
|
|
$ |
4,557,233.61 |
|
|
|
Invus Public Equities, L.P.(2)
|
|
|
|
|
2,492,646 |
|
|
|
|
$ |
24,999,993.06 |
|
|
|
|
$ |
— |
|
|
|
Entities affiliated with Fidelity(3)
|
|
|
|
|
2,492,646 |
|
|
|
|
$ |
24,999,993.07 |
|
|
|
|
$ |
— |
|
|
|
John Maraganore, Ph.D.(4)
|
|
|
|
|
56,095 |
|
|
|
|
$ |
— |
|
|
|
|
$ |
506,359.29 |
|
|
(1)
Robert Nelsen, M.B.A. serves as a member of our board of directors and is an affiliate of ARCH Venture Partners, of which Venture Fund XII, L.P. is an affiliated fund. Venture Fund XII, L.P. beneficially owns more than 5% of our outstanding capital stock.
(2)
Invus Public Equities, L.P. beneficially owns more than 5% of our outstanding capital stock.
(3)
Fidelity collectively refers to Powhatan & Co., LLC FBO Fidelity Growth Company Private Investments Portfolio LLC; Fidelity Mt. Vernon Street Trust: Fidelity Growth Company Fund; Fidelity Mt. Vernon Street Trust: Fidelity Growth Company K6 Fund; and Fidelity Mt. Vernon Street Trust: Fidelity Series Growth Company Fund. Fidelity beneficially owns more than 5% of our outstanding capital stock.
(4)
John Maraganore, Ph.D., is our co-founder and Executive Chair, and holds more than 5% of our outstanding capital stock.
Series B Convertible Preferred Stock Financing
On May 20, 2026 and June 4, 2026, we issued and sold an aggregate of 6,801,318 shares of our Series B convertible preferred stock, par value $0.0001 per share, at a purchase price of $14.6295 per share, for an aggregate purchase price of approximately $99.5 million.
The outstanding shares of Series B convertible preferred stock will convert into shares of our common stock at a rate of one share of common stock immediately prior to the completion of this offering. The following table summarizes the shares of our Series B convertible preferred stock issued to our related parties:
|
Purchasers
|
|
|
Shares of Series B
Convertible
Preferred Stock
|
|
|
Total Purchase
Price
|
|
|
ARCH Venture Fund XII, L.P.(1)
|
|
|
|
|
1,845,585 |
|
|
|
|
$ |
26,999,985.76 |
|
|
|
Invus Public Equities, L.P.(2)
|
|
|
|
|
446,320 |
|
|
|
|
$ |
6,529,438.44 |
|
|
|
Funds and/or accounts advised by affiliates of FMR LLC(3)
|
|
|
|
|
1,025,325 |
|
|
|
|
$ |
14,999,992.09 |
|
|
|
Entities affiliated with Viking(4)
|
|
|
|
|
1,845,585 |
|
|
|
|
$ |
26,999,985.70 |
|
|
(1)
Robert Nelsen, M.B.A. serves as a member of our board of directors and is an affiliate of ARCH Venture Partners, of which Venture Fund XII, L.P. is an affiliated fund. Venture Fund XII, L.P. beneficially owns more than 5% of our outstanding capital stock.
(2)
Invus Public Equities, L.P. beneficially owns more than 5% of our outstanding capital stock.
(3)
Reflects shares held by Fidelity Advisor Series VII: Fidelity Advisor Biotechnology Fund; Powhatan & Co., LLC FBO Fidelity Growth Company Private Investments Portfolio LLC; Fidelity Mt. Vernon Street Trust: Fidelity Growth Company Fund; Fidelity Mt. Vernon Street Trust: Fidelity Growth Company K6 Fund; Fidelity Mt. Vernon Street Trust: Fidelity Series Growth Company Fund; Fidelity Select Portfolios: Biotechnology Portfolio and Powhatan & Co., LLC FBO Fidelity Growth Company Private Investments Portfolio LLC (collectively, “FMR LLC Entities”). FMR LLC, as adviser to the FMR LLC Entities, beneficially owns or is deemed to beneficially own more than 5% of our outstanding capital stock.
(4)
Viking collectively refers to Viking Global Opportunities Illiquid Investment Sub-Master LP and Viking Global Opportunities Vintage IV (Aggregator) LP. Viking beneficially owns more than 5% of our outstanding capital stock.
Maraganore Consulting Agreement
On August 14, 2023, we entered into a consulting agreement (the “Maraganore Consulting Agreement”) with Dr. Maraganore, pursuant to which we granted Dr. Maraganore 823,725 shares of our common stock at the fair market value per share on the grant date, subject to certain vesting conditions. Under the Maraganore Consulting Agreement, Dr. Maraganore provides certain strategic advisory services to the Company. The Maraganore Consulting Agreement remains in effect unless earlier terminated or extended by the parties. In connection with the completion of this offering, we intend to terminate the Maraganore Consulting Agreement. Dr. Maraganore will continue to serve as the Executive Chair of our board of directors. Following the termination of the Maraganore Consulting Agreement, Dr. Maraganore will be eligible to receive compensation for his role as Executive Chair of our board of directors pursuant to our non-employee director compensation policy, which will become effective upon the effectiveness of the registration statement of which this prospectus forms a part. See “Executive Compensation — Consulting Agreement with Dr. Maraganore.”
Maraganore Executive Chair Agreement
On November 10, 2023, we entered into an executive chair agreement (as amended, the “Maraganore Executive Chair Agreement”) with Dr. Maraganore, pursuant to which Dr. Maraganore agreed to provide services as Executive Chair and a member of our board of directors, and we agreed
to pay Dr. Maraganore an annual cash retainer of $200,000, in addition to the grant of options to purchase an aggregate of 471,256 shares of our common stock, with 25% of the shares subject to each option vesting after 12 months of continuous service from the applicable vesting commencement date and the remaining shares vesting in equal monthly installments over the following 36 months of continuous service. In the event of an Involuntary Termination, as defined in the Maraganore Executive Chair Agreement, all remaining unvested shares subject to the options will immediately vest.
In addition, the Maraganore Executive Chair Agreement provides that Dr. Maraganore is eligible to be considered for additional equity grants at the discretion of our board of directors. In May and June 2024, Dr. Maraganore received performance-based options to purchase an aggregate of 201,966 shares of our common stock, which subsequently vested upon the achievement of a specified regulatory development milestone. Additionally, in February and June 2026, Dr. Maraganore received options to purchase an aggregate of 340,553 shares of our common stock, consisting of 135,611 shares that vest monthly over 48 months of continuous service and two tranches of 102,471 shares of our common stock that vest upon the achievement of specified clinical and regulatory development milestones, respectively, provided that each applicable milestone is achieved within 10 years following the applicable date of grant.
The Maraganore Executive Chair Agreement will continue until Dr. Maraganore ceases to be a member of our board of directors, which may occur at any time at the election of Dr. Maraganore or the Company. In the event that the Company successfully consummates this offering on or prior to December 31, 2026, the annual cash retainer for Dr. Maraganore will increase to $240,000, effective as of the consummation of this offering. During each of the years ended December 31, 2024 and 2025, we paid Dr. Maraganore approximately $0.2 million for services provided under the Maraganore Executive Chair Agreement.
Management Rights and Side Letters
In connection with our convertible preferred stock financings, we entered into management rights and side letters with certain purchasers of our convertible preferred stock, including holders of more than 5% of our capital stock and entities with which certain of our directors are affiliated, pursuant to which such entities were granted certain management rights, among other things, including the right to consult with and advise our management on significant business issues, review our operating plans, examine our books, and records and inspect our facilities. Except for certain confidentiality obligations, publicity restrictions, and market standoff provisions, the rights under these management rights and side letters will terminate immediately prior to the completion of this offering.
Agreements with our Stockholders
In connection with our convertible preferred stock financings, we entered into an investors’ rights agreement, a right of first refusal and co-sale agreement, and voting agreement, in each case, with the purchasers of our convertible preferred stock. Our amended and restated right of first refusal and co-sale agreement (“ROFR Agreement”) provides for rights of first refusal and co-sale and drag along rights in respect of sales by certain holders of our capital stock. Our amended and restated voting agreement, as amended (“Voting Agreement”), contains provisions with respect to the election of our board of directors and its composition. The rights under each of the ROFR Agreement and Voting Agreement will terminate upon the closing of this offering.
Our amended and restated investors’ rights agreement (“Investor Rights Agreement”) provides certain holders of our convertible preferred stock with a participation right to purchase their pro rata share of new securities that we may propose to sell and issue, subject to certain exceptions. Such participation right will terminate upon the closing of this offering. The Investor Rights Agreement further provides certain holders of our capital stock with the right to demand that we file a registration statement, subject to certain limitations, and to request that their shares be covered by a registration statement that we are otherwise filing. See the section titled “Description of Capital Stock — Registration Rights” appearing elsewhere in this prospectus, for additional information regarding such registration rights.
Employment Arrangements
We have entered into offer letter agreements with certain of our executive officers and granted stock options to our executive officers, as more fully described in the section titled “Executive Compensation.”
Equity Grants
We have granted stock options to purchase our shares of common stock to certain of our executive officers and directors. For more information regarding the stock options granted to our executive officers and directors, see the sections titled “Executive Compensation” and “Director Compensation” included elsewhere in this prospectus.
Indemnification Agreements
Our amended and restated certificate of incorporation will contain provisions limiting the liability of directors and officers, and our amended and restated bylaws will provide that we will indemnify each of our directors and officers to the fullest extent permitted under Delaware law. Our amended and restated certificate of incorporation and amended and restated bylaws will also provide our board of directors with discretion to indemnify our employees and other agents when determined appropriate by the board of directors. In addition, we have entered into or intend to enter into an indemnification agreement with each of our directors and executive officers, which will require us to indemnify them. For more information regarding these agreements, see the section titled “Management — Limitations on Liability and Indemnification.”
Policies and Procedures for Transactions with Related Persons
In connection with this offering, we have adopted a written policy, to become effective upon the effectiveness of the registration statement of which this prospectus forms a part, that our executive officers, directors, nominees for election as a director, beneficial owners of more than 5% of any series of our common stock, and any members of the immediate family of any of the foregoing persons are not permitted to enter into a related person transaction with us without the approval or ratification of our board of directors or our audit committee. Any request for us to enter into a transaction with an executive officer, director, nominee for election as a director, beneficial owner of more than 5% of any series of our common stock, or any member of the immediate family of any of the foregoing persons, in which the amount involved exceeds $120,000 (or, if less, 1% of the average of our total assets in a fiscal year) and such person would have a direct or indirect interest, must be presented to our board of directors or our audit committee for review, consideration, and approval. In approving or rejecting any such proposal, our board of directors or our audit committee is to consider the material facts of the transaction, including whether the transaction is on terms no less favorable than terms generally available to an unaffiliated third party under the same or similar circumstances and the extent of the related person’s interest in the transaction.
PRINCIPAL STOCKHOLDERS
The following table sets forth information regarding beneficial ownership of our capital stock as of September 11, 2026 by:
•
each person, or group of affiliated persons, known by us to beneficially own more than 5% of our common stock;
•
each of our directors and director nominees;
•
each of our named executive officers; and
•
all of our current executive officers and directors as a group.
We have determined beneficial ownership in accordance with the rules of the SEC. Under these rules, beneficial ownership includes any shares of common stock as to which the individual or entity has sole or shared voting power or investment power. Unless otherwise indicated below, to our knowledge the persons and entities named in the table have sole voting and sole investment power with respect to all shares that they beneficially owned, subject to community property laws where applicable. We have deemed shares of common stock subject to options that are currently exercisable or exercisable within 60 days of September 11, 2026 to be outstanding and to be beneficially owned by the person holding the option for the purpose of computing the percentage ownership of that person but have not treated them as outstanding for the purpose of computing the percentage ownership of any other person.
Applicable percentage ownership before the offering is based on an aggregate of 38,675,212 shares of common stock (which includes 1,820,501 shares of restricted common stock subject to repurchase or forfeiture) deemed to be outstanding as of September 11, 2026, after giving effect to the automatic conversion of all outstanding shares of convertible preferred stock into shares of common stock immediately prior to the completion of this offering, and assuming an initial public offering price of $18.00 per share, which is the midpoint of the estimated offering price range set forth on the cover page of this prospectus.
Applicable percentage ownership after the offering is based on 50,562,595 shares of common stock assumed to be outstanding immediately after the completion of this offering (including the sale of shares of common stock in this offering and assuming no exercise of the underwriters’ option to purchase additional shares).
Unless otherwise indicated, the address for each beneficial owner listed in the table below is c/o City Therapeutics, Inc., 399 Binney Street, Cambridge, Massachusetts 02142.
| |
|
|
Shares of
Common
Stock
Beneficially
Owned
|
|
|
Percentage of Shares
Beneficially Owned
|
|
|
Name of Beneficial Owner
|
|
|
Before
Offering
|
|
|
After
Offering
|
|
|
5% or Greater Stockholders:
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
ARCH Venture Fund XII, L.P.(1)
|
|
|
|
|
13,522,507 |
|
|
|
|
|
34.96% |
|
|
|
|
|
26.74% |
|
|
|
Invus Public Equities, L.P.(2)
|
|
|
|
|
3,227,866 |
|
|
|
|
|
8.35% |
|
|
|
|
|
6.38% |
|
|
|
FMR LLC(3)
|
|
|
|
|
3,863,783 |
|
|
|
|
|
9.99% |
|
|
|
|
|
7.64% |
|
|
|
Entities Affiliated with Viking(4)
|
|
|
|
|
2,027,005 |
|
|
|
|
|
5.24% |
|
|
|
|
|
4.01% |
|
|
|
Named Executive Officers and Directors:
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
Andrew (Andy) Orth, M.B.A.(5)
President, Chief Executive Officer and Director
|
|
|
|
|
1,094,544 |
|
|
|
|
|
2.83% |
|
|
|
|
|
2.16% |
|
|
|
Baisong Mei, M.D., Ph.D.(6)
Chief Medical Officer
|
|
|
|
|
234,678 |
|
|
|
|
|
* |
|
|
|
|
|
* |
|
|
|
Tracy Zimmermann, Ph.D.(7)
Chief Scientific Officer
|
|
|
|
|
269,569 |
|
|
|
|
|
* |
|
|
|
|
|
* |
|
|
|
John Maraganore, Ph.D.(8)
|
|
|
|
|
2,121,831 |
|
|
|
|
|
5.48% |
|
|
|
|
|
4.20% |
|
|
|
Barry Greene(9)
|
|
|
|
|
31,186 |
|
|
|
|
|
* |
|
|
|
|
|
* |
|
|
|
Robert Nelsen, M.B.A.(1)
|
|
|
|
|
— |
|
|
|
|
|
* |
|
|
|
|
|
* |
|
|
|
Saraswathy (Sara) V. Nochur, Ph.D.(10)
|
|
|
|
|
27,443 |
|
|
|
|
|
* |
|
|
|
|
|
* |
|
|
|
Ron Philip(11)
|
|
|
|
|
33,680 |
|
|
|
|
|
* |
|
|
|
|
|
* |
|
|
|
All executive officers and directors as a group (10 persons)(12)
|
|
|
|
|
4,355,351 |
|
|
|
|
|
11.14% |
|
|
|
|
|
8.54% |
|
|
*
Represents beneficial ownership of less than 1%.
(1)
Consists of (i) 2,718,292 shares of common stock, (ii) 3,294,900 shares of common stock issuable upon conversion of Series Seed convertible preferred stock, (iii) 5,482,309 shares of common stock issuable upon conversion of Series A convertible preferred stock, and (iv) 2,027,006 shares of common stock issuable upon conversion of Series B convertible preferred stock held by ARCH Venture Fund XII, L.P. (“ARCH XII”). ARCH Venture Partners XII, L.P. (“AVP XII LP”) is the general partner of ARCH XII. ARCH Venture Partners XII, LLC (“AVP XII LLC”) is the general partner of AVP XII LP. Keith Crandell, Kristina Burow, Steven Gillis and Robert Nelsen, a member of our board of directors, comprise the investment committee of AVP XII LLC (the “AVP XII LLC Committee Members”). Each of AVP XII LP and AVP XII LLC may be deemed to beneficially own the shares held by ARCH XII, and each of the AVP XII LLC Committee Members may be deemed to share the power to direct the disposition and vote of the shares held by ARCH XII. Each of AVP XII LP, AVP XII LLC and the AVP XII LLC Committee Members disclaims beneficial ownership except to the extent of their pecuniary interest therein, if any. The address for ARCH Venture Partners and the individuals listed above is c/o ARCH Venture Partners, 8755 W. Higgins Rd., Suite 1025, Chicago, IL 60631.
(2)
Consists of (i) 2,737,673 shares of common stock issuable upon conversion of Series A convertible preferred stock, and (ii) 490,193 shares of common stock issuable upon conversion of Series B convertible preferred stock held by Invus Public Equities, L.P. (“IPE”). Invus Public Equities Advisors, LLC (“IPEA”) controls IPE, as its general partner and accordingly, may be deemed to beneficially own the shares held by IPE. Invus Global Management, LLC (“IGM”) controls IPEA, as its managing member and accordingly, may be deemed to beneficially own the shares that IPEA may be deemed to beneficially own. Siren, L.L.C. (“Siren”) controls IGM, as its managing member and accordingly, may be deemed to beneficially own the shares that IGM may be deemed to beneficially own. Raymond Debbane, as the managing member of Siren, controls Siren and accordingly, may be deemed to beneficially own the shares that Siren may be deemed to beneficially own. The business address for IPE, IPEA, IGM, Siren and Mr. Raymond Debbane is 750 Lexington Ave, 30th Floor, New York, New York 10022.
(3)
Consists of (i) 1,314,166 shares of common stock issuable upon the conversion of Series A convertible preferred stock held by Powhatan & Co., LLC FBO Fidelity Growth Company Private Investments Portfolio LLC, (ii) 879,695 shares of common stock issuable upon the conversion of Series A convertible preferred stock held by Fidelity Mt. Vernon Street Trust: Fidelity Growth Company Fund, (iii) 328,267 shares of common stock issuable upon the conversion of Series A convertible preferred stock held by Fidelity Mt. Vernon Street Trust: Fidelity Growth Company K6 Fund, (iv) 215,543 shares of common stock issuable upon the conversion of Series A convertible preferred stock held by Fidelity Mt. Vernon Street Trust: Fidelity Series Growth Company Fund, (v) 47,446 shares of common stock issuable upon the conversion of Series B convertible preferred stock held by Fidelity Advisor Series VII: Fidelity Advisor Biotechnology Fund, (vi) 151,675 shares of common stock issuable upon the conversion of Series B convertible preferred stock held by Powhatan & Co., LLC FBO Fidelity Growth Company Private Investments Portfolio LLC, (vii) 114,992 shares of common stock issuable upon the conversion of Series B convertible preferred stock held by Fidelity Mt. Vernon Street Trust: Fidelity Growth Company Fund, (viii) 32,070 shares of
common stock issuable upon the conversion of Series B convertible preferred stock held by Fidelity Mt. Vernon Street Trust: Fidelity Growth Company K6 Fund, (ix) 28,363 shares of common stock issuable upon the conversion of Series B convertible preferred stock held by Fidelity Mt. Vernon Street Trust: Fidelity Series Growth Company Fund, and (x) 751,566 shares of common stock issuable upon the conversion of Series B convertible preferred stock held by Fidelity Select Portfolios: Biotechnology Portfolio. The funds and accounts identified in this footnote are advised or managed by direct or indirect subsidiaries of FMR LLC, and the shares held by such funds or accounts are beneficially owned, or may be deemed to be beneficially owned, by FMR LLC, certain of its subsidiaries, affiliates, and other companies. Abigail P. Johnson is a director, the Chairman, and the Chief Executive Officer of FMR LLC. Members of the Johnson family including Abigail P. Johnson, are the predominant owners, directly or through trusts, of Series B voting common shares representing 49% of the voting power of FMR LLC. The Johnson family group and all other Series B shareholders have entered into a shareholders’ voting agreement under which all Series B voting common shares will be voted in accordance with the majority vote of Series B voting common shares. Accordingly, through their ownership of voting common shares and the execution of the shareholders’ voting agreement, members of the Johnson family may be deemed, under the Investment Company Act of 1940, to form a controlling group with respect to FMR LLC. The address for FMR LLC is 245 Summer Street, Boston, MA 02210.
(4)
Consists of (i) 1,418,904 shares of common stock issuable upon the conversion of Series B convertible preferred stock held by Viking Global Opportunities Illiquid Investments Sub-Master LP (the “Opportunities Fund”) and (ii) 608,101 shares of common stock issuable upon the conversion of Series B convertible preferred stock held by Viking Global Opportunities Vintage IV (Aggregator) LP (the “Vintage IV Fund”). The Opportunities Fund has the authority to dispose of and vote the shares directly owned by it, which power may be exercised by its general partner, Viking Global Opportunities Portfolio GP LLC (the “Opportunities GP”), and by Viking Global Investors LP (“VGI”), which provides managerial services to the Opportunities Fund. The Vintage IV Fund has the authority to dispose of and vote the shares directly owned by it, which power may be exercised by its general partner, Viking Global Opportunities Vintage IV Portfolio GP LLC (the “Vintage IV GP”) and VGI, which provides managerial services to the Vintage IV Fund. O. Andreas Halvorsen and Rose S. Shabet, as Executive Committee Members of Viking Global Partners LLC (the general partner of VGI) and Viking Global Opportunities Parent GP LLC (the “Parent GP”), the sole member of each of (i) Viking Global Opportunities GP LLC (which is the sole member of the Opportunities GP) and (ii) Viking Global Opportunities Vintage IV GP LLC (which is the sole member of the Vintage IV GP), have shared authority to dispose of and vote the shares of common stock beneficially owned by VGI and the Parent GP and each may be deemed to beneficially own the shares of common stock that the Opportunities Fund and the Vintage IV Fund directly own. The business address of the entities is c/o Viking Global Investors LP, 600 Washington Boulevard, Floor 11, Stamford, Connecticut 06901.
(5)
Consists of (i) 1,063,154 shares of common stock (including restricted common stock subject to repurchase or forfeiture) and (ii) 31,390 shares of common stock underlying outstanding stock options that are immediately exercisable within 60 days of September 11, 2026.
(6)
Consists of (i) 228,743 shares of common stock (including restricted common stock subject to repurchase or forfeiture) and (ii) 5,935 shares of common stock underlying outstanding stock options that are immediately exercisable within 60 days of September 11, 2026.
(7)
Consists of 269,569 shares of common stock underlying outstanding stock options that are immediately exercisable within 60 days of September 11, 2026.
(8)
Consists of: (i) 859,933 shares of common stock (including restricted common stock subject to repurchase or forfeiture) held by Dr. Maraganore, (ii) 549,150 shares of common stock issuable upon the conversion of Series Seed convertible preferred stock, (iii) 61,609 shares of common stock issuable upon the conversion of Series A convertible preferred stock, (iv) 318,507 shares of common stock held by the John M. Maraganore 2021 Irrevocable Trust, (v) 318,507 shares of common stock held by the Christine Maraganore 2021 Irrevocable Trust, and (vi) 14,125 shares of common stock underlying outstanding stock options that are immediately exercisable within 60 days of September 11, 2026. Dr. Maraganore may be deemed to beneficially own the shares held by the John M. Maraganore 2021 Irrevocable Trust and the Christine Maraganore 2021 Irrevocable Trust.
(9)
Consists of 31,186 shares of common stock underlying outstanding stock options that are immediately exercisable within 60 days of September 11, 2026.
(10)
Consists of 27,443 shares of common stock underlying outstanding stock options that are immediately exercisable within 60 days of September 11, 2026.
(11)
Consists of 33,680 shares of common stock underlying outstanding stock options that are immediately exercisable within 60 days of September 11, 2026.
(12)
Consists of (i) 3,323,355 shares of common stock (including restricted common stock subject to repurchase or forfeiture), (ii) 549,150 shares of common stock issuable upon the conversion of Series Seed convertible preferred stock, (iii) 61,609 shares of common stock issuable upon the conversion of Series A convertible preferred stock, and (iv) 421,237 shares of common stock underlying outstanding stock options that are immediately exercisable within 60 days of September 11, 2026.
DESCRIPTION OF CAPITAL STOCK
General
The following description of our capital stock and certain provisions of our amended and restated certificate of incorporation and amended and restated bylaws are summaries and are qualified by reference to the amended and restated certificate of incorporation, which will become effective immediately prior to the completion of this offering, and the amended and restated bylaws, which will become effective upon the effectiveness of the registration statement of which this prospectus forms a part. Copies of these documents have been filed with the SEC as exhibits to our registration statement, of which this prospectus forms a part. The descriptions of the common stock and preferred stock reflect changes to our capital structure that will be in effect on the completion of this offering.
Upon the filing of our amended and restated certificate of incorporation and the completion of this offering, our authorized capital stock will consist of 500,000,000 shares of common stock, par value $0.0001 per share, and 10,000,000 shares of preferred stock, par value $0.0001 per share, all of which shares of preferred stock will be undesignated.
As of June 30, 2026, there were 38,476,269 shares of common stock (which includes 1,964,421 shares of restricted common stock subject to repurchase or forfeiture) outstanding and held of record by stockholders. This amount assumes the conversion of all outstanding shares of our convertible preferred stock into common stock, which will occur immediately prior to the completion of this offering.
Common Stock
Holders of our common stock are entitled to one vote for each share held on all matters submitted to a vote of the stockholders. The holders of our common stock do not have any cumulative voting rights. Holders of our common stock are entitled to receive ratably any dividends declared by our board of directors out of funds legally available for that purpose, subject to any preferential dividend rights of any outstanding convertible preferred stock. Our common stock has no preemptive rights, conversion rights or other subscription rights or redemption or sinking fund provisions.
In the event of our liquidation, dissolution or winding up, holders of our common stock will be entitled to share ratably in all assets remaining after payment of all debts and other liabilities and any liquidation preference of any outstanding preferred stock. The shares to be issued by us in this offering will be, when issued and paid for, validly issued, fully paid and non-assessable.
Preferred Stock
Immediately prior to the completion of this offering, all outstanding shares of our convertible preferred stock will be converted into shares of our common stock. Upon the consummation of this offering, our board of directors will have the authority, without further action by our stockholders, to issue up to 10,000,000 shares of preferred stock in one or more series and to fix the rights, preferences, privileges and restrictions thereof. These rights, preferences and privileges could include dividend rights, conversion rights, voting rights, terms of redemption, liquidation preferences, sinking fund terms and the number of shares constituting, or the designation of, such series, any or all of which may be greater than the rights of common stock. The issuance of our preferred stock could adversely affect the voting power of holders of common stock and the likelihood that such holders will receive dividend payments and payments upon our liquidation. In addition, the issuance of preferred stock could have the effect of delaying, deferring or preventing a change in control of our company or other corporate action. Immediately after consummation of this offering, no shares of preferred stock will be outstanding, and we have no present plan to issue any shares of preferred stock.
Stock Options
As of June 30, 2026, 5,404,513 shares of common stock were issuable upon the exercise of outstanding stock options under the 2023 Plan, at a weighted-average exercise price of $1.97 per share; and shares of our common stock were reserved for future issuance under the 2026 Plan, which
will become effective once the registration statement of which this prospectus forms a part is declared effective, as well as any future automatic annual increases in the number of shares of common stock reserved for issuance under the 2026 Plan, and any shares underlying outstanding stock awards granted under the 2023 Plan, that expire or are repurchased, forfeited, cancelled or withheld. For additional information regarding terms of our equity incentive plans, see the section titled “Executive Compensation — Employee Benefit and Equity Compensation Plans.”
Registration Rights
Upon the completion of this offering and subject to the lock-up agreements entered into in connection with this offering and federal securities laws, certain holders of shares of our common stock, including those shares of our common stock that will be issued upon the conversion of our convertible preferred stock in connection with this offering, will initially be entitled to certain rights with respect to registration of such shares under the Securities Act. These shares are referred to as registrable securities. The holders of these registrable securities possess registration rights pursuant to the terms of the Investor Rights Agreement and are described in additional detail below. The registration of shares of our common stock pursuant to the exercise of the registration rights described below would enable the holders to trade these shares without restriction under the Securities Act when the applicable registration statement is declared effective. We will pay all registration expenses, other than underwriting discounts, selling commissions and stock transfer taxes, of the shares registered pursuant to the demand, piggyback and Form S-3 registrations described below.
Generally, in an underwritten offering, the managing underwriter, if any, has the right, subject to specified conditions and limitations, to limit the number of shares the holders may include. The demand, piggyback and Form S-3 registration rights described below will expire no later than four years after the completion of this offering.
Form S-1 Registration Rights
Upon the completion of this offering, certain holders of our common stock, including those issuable upon the conversion of shares of our convertible preferred stock upon completion of this offering, will be entitled to certain demand registration rights. At any time beginning 180 days after the completion of this offering, the holders of a majority of registrable securities then outstanding may request that we register all or a portion of their shares on Form S-1 if the anticipated aggregate offering price of the shares offered would exceed $75 million. With certain exceptions, we are not required to effect the filing of a registration statement during the period starting with the date of the filing of, and ending on a date 60 days following the effective date of the registration statement for this offering. We will not be required to effect more than two registrations pursuant to this provision of the Investor Rights Agreement.
Piggyback Registration Rights
In connection with this offering, certain holders of our common stock, including those issuable upon the conversion of shares of our convertible preferred stock upon completion of this offering, were entitled to, and the necessary percentage of holders waived, their rights to notice of this offering and to include their shares of registrable securities in this offering. After this offering, in the event that we propose to register any of our securities under the Securities Act, either for our own account or for the account of other security holders, the holders of these shares will be entitled to certain piggyback registration rights allowing the holder to include their shares in such registration, subject to certain marketing and other limitations.
Form S-3 Registration Rights
Upon the completion of this offering, certain holders of our common stock, including those issuable upon the conversion of shares of our convertible preferred stock upon completion of this offering, will be entitled to certain Form S-3 registration rights. Holders of at least 20% of registrable securities then outstanding can make a request that we register their shares on Form S-3 if we are qualified to file a registration statement on Form S-3 and if the anticipated aggregate offering price of the shares offered would exceed $5 million. We will not be required to effect more than two registrations on Form S-3 within
any twelve-month period pursuant to this provision of the Investor Rights Agreement. The right to have such shares registered on Form S-3 is further subject to other specified conditions and limitations.
Expiration of Registration Rights
The demand registration rights and short-form registration rights granted under the investor rights agreement will terminate on the earliest of (i) the closing of a “Deemed Liquidation Event,” as such term is defined in our certificate of incorporation (as currently in effect), (ii) the fifth anniversary of the completion of this offering and (iii) such time after consummation of this offering as SEC Rule 144 or another similar exemption under the Securities Act is available for the sale of all of the securities subject to such registration rights without limitation during a three-month period without registration.
Anti-Takeover Effects of Our Certificate of Incorporation and Bylaws and Delaware Law
Our amended and restated certificate of incorporation and amended and restated bylaws will include a number of provisions that may have the effect of delaying, deferring or preventing another party from acquiring control of us and encouraging persons considering unsolicited tender offers or other unilateral takeover proposals to negotiate with our board of directors rather than pursue non-negotiated takeover attempts. These provisions include the items described below.
Board Composition and Filling Vacancies
Our amended and restated certificate of incorporation will provide for the division of our board of directors into three classes serving staggered three-year terms, with one class being elected each year. Our amended and restated certificate of incorporation also will provide that directors may be removed only for cause and then only by the affirmative vote of the holders of two-thirds or more of the shares then entitled to vote at an election of directors. Furthermore, any vacancy on our board of directors, however occurring, including a vacancy resulting from an increase in the size of our board, may only be filled by the affirmative vote of a majority of our directors then in office even if less than a quorum. The classification of directors, together with the limitations on removal of directors and treatment of vacancies, has the effect of making it more difficult for stockholders to change the composition of our board of directors.
No Written Consent of Stockholders
Our amended and restated certificate of incorporation will provide that all stockholder actions are required to be taken by a vote of the stockholders at an annual or special meeting, and that stockholders may not take any action by written consent in lieu of a meeting. This limit may lengthen the amount of time required to take stockholder actions and would prevent the amendment of our amended and restated bylaws or removal of directors by our stockholders without holding a meeting of stockholders.
Meetings of Stockholders
Our amended and restated certificate of incorporation and amended and restated bylaws will provide that only a majority of the members of our board of directors then in office may call special meetings of stockholders and only those matters set forth in the notice of the special meeting may be considered or acted upon at a special meeting of stockholders. Our amended and restated bylaws limit the business that may be conducted at an annual meeting of stockholders to those matters properly brought before the meeting.
Advance Notice Requirements
Our amended and restated bylaws will establish advance notice procedures with regard to stockholder proposals relating to the nomination of candidates for election as directors or new business to be brought before meetings of our stockholders. These procedures provide that notice of stockholder proposals must be timely given in writing to our corporate secretary prior to the meeting at which the action is to be taken. Generally, to be timely, notice must be received at our principal executive offices not less than 90 days nor more than 120 days prior to the first anniversary date of the annual meeting for
the preceding year. Our amended and restated bylaws specify the requirements as to form and content of all stockholders’ notices. These requirements may preclude stockholders from bringing matters before the stockholders at an annual or special meeting.
Amendment to Certificate of Incorporation and Bylaws
Any amendment to our amended and restated certificate of incorporation must first be approved by a majority of our board of directors, and if required by law or our amended and restated certificate of incorporation, must thereafter be approved by a majority of the outstanding shares entitled to vote on the amendment and a majority of the outstanding shares of each class entitled to vote thereon as a class, except that the amendment of the provisions relating to stockholder action, board composition and limitation of liability must be approved by not less than two-thirds of the outstanding shares entitled to vote on the amendment, and not less than two-thirds of the outstanding shares of each class entitled to vote thereon as a class. Our amended and restated bylaws may be amended by the affirmative vote of a majority of the directors then in office, subject to any limitations set forth in the amended and restated bylaws; and may also be amended by the affirmative vote of a majority of the outstanding shares entitled to vote on the amendment, voting together as a single class, except that the amendment of the provisions relating to notice of stockholder business and nominations and special meetings must be approved by not less than two-thirds of the outstanding shares entitled to vote on the amendment, and not less than two-thirds of the outstanding shares of each class entitled to vote thereon as a class, or, if our board of directors recommends that the stockholders approve the amendment, by the affirmative vote of the majority of the outstanding shares entitled to vote on the amendment, in each case voting together as a single class.
Undesignated Preferred Stock
Our amended and restated certificate of incorporation will provide for 10,000,000 authorized shares of preferred stock. The existence of authorized but unissued shares of preferred stock may enable our board of directors to discourage an attempt to obtain control of us by means of a merger, tender offer, proxy contest or otherwise. For example, if in the due exercise of its fiduciary obligations, our board of directors were to determine that a takeover proposal is not in the best interests of our stockholders, our board of directors could cause shares of preferred stock to be issued without stockholder approval in one or more private offerings or other transactions that might dilute the voting or other rights of the proposed acquirer or insurgent stockholder or stockholder group. In this regard, our amended and restated certificate of incorporation will grant our board of directors broad power to establish the rights and preferences of authorized and unissued shares of preferred stock. The issuance of shares of preferred stock could decrease the amount of earnings and assets available for distribution to holders of shares of common stock. The issuance may also adversely affect the rights and powers, including voting rights, of these holders and may have the effect of delaying, deterring or preventing a change in control of us.
Section 203 of the Delaware General Corporation Law
Upon completion of this offering, we will be subject to the provisions of Section 203 of the DGCL. In general, Section 203 prohibits a publicly held Delaware corporation from engaging in a “business combination” with an “interested stockholder” for a three-year period following the time that this stockholder becomes an interested stockholder, unless the business combination is approved in a prescribed manner. Under Section 203, a business combination between a corporation and an interested stockholder is prohibited unless it satisfies one of the following conditions:
•
before the stockholder became interested, our board of directors approved either the business combination or the transaction which resulted in the stockholder becoming an interested stockholder;
•
upon consummation of the transaction which resulted in the stockholder becoming an interested stockholder, the interested stockholder owned at least 85% of the voting stock of the corporation outstanding at the time the transaction commenced, excluding for purposes of determining the voting stock outstanding, shares owned by persons who are directors and
also officers, and employee stock plans, in some instances, but not the outstanding voting stock owned by the interested stockholder; or
•
at or after the time the stockholder became interested, the business combination was approved by our board of directors and authorized at an annual or special meeting of the stockholders by the affirmative vote of at least two-thirds of the outstanding voting stock which is not owned by the interested stockholder.
Section 203 defines a business combination to include:
•
any merger or consolidation involving the corporation and the interested stockholder;
•
any sale, transfer, lease, pledge or other disposition involving the interested stockholder of 10% or more of the assets of the corporation;
•
subject to exceptions, any transaction that results in the issuance or transfer by the corporation of any stock of the corporation to the interested stockholder;
•
subject to exceptions, any transaction involving the corporation that has the effect of increasing the proportionate share of the stock of any class or series of the corporation beneficially owned by the interested stockholder; and
•
the receipt by the interested stockholder of the benefit of any loans, advances, guarantees, pledges or other financial benefits provided by or through the corporation.
In general, Section 203 defines an interested stockholder as any entity or person beneficially owning 15% or more of the outstanding voting stock of the corporation and any entity or person affiliated with or controlling or controlled by the entity or person.
Choice of Forum
Our amended and restated bylaws will provide that the Court of Chancery of the State of Delaware is the sole and exclusive forum for the following claims or causes of action under the Delaware statutory or common law: (i) any derivative action or proceeding brought on our behalf, (ii) any action asserting a claim of, or a claim based on, a breach of a fiduciary duty owed by any of our current or former directors, officers or other employees or stockholders to us or our stockholders, (iii) any action asserting a claim arising pursuant to any provision of the DGCL or our amended and restated certificate of incorporation or amended and restated bylaws (including the interpretation, validity or enforceability thereof) or as to which the DGCL confers jurisdiction on the Court of Chancery of the State of Delaware or (iv) any action asserting a claim governed by the internal affairs doctrine.
However, Section 27 of the Exchange Act creates exclusive federal jurisdiction over all claims brought to enforce any duty or liability created by the Exchange Act or the rules and regulations thereunder. Consequently, this choice of forum provision would not apply to claims or causes of action brought to enforce a duty or liability created by the Exchange Act or any other claim for which the federal courts have exclusive jurisdiction or the Securities Act. Moreover, Section 22 of the Securities Act creates concurrent jurisdiction for federal and state courts over all claims brought to enforce any duty or liability created by the Securities Act or the rules and regulations thereunder.
In addition, our amended and restated bylaws will provide that, unless we consent in writing to the selection of an alternative forum, to the fullest extent permitted by law, the federal district courts of the United States of America shall be the exclusive forum for the resolution of any complaint asserting a cause or causes of action arising under the Securities Act, including all causes of action asserted against any defendant to such complaint. For the avoidance of doubt, this provision is intended to benefit and may be enforced by us, our officers and directors, the underwriters to any offering giving rise to such complaint, and any other professional entity whose profession gives authority to a statement made by that person or entity and who has prepared or certified any part of the documents underlying the offering.
While the Delaware courts have determined that such choice of forum provisions are facially valid, a stockholder may nevertheless seek to bring a claim in a venue other than those designated in the
exclusive forum provisions, and there can be no assurance that such provisions will be enforced by a court in those other jurisdictions. We note that investors cannot waive compliance with the federal securities laws and the rules and regulations thereunder.
Additionally, our amended and restated bylaws will provide that any person or entity holding, owning or otherwise acquiring any interest in any of our securities shall be deemed to have notice of and consented to these provisions.
Limitations on Liability and Indemnification
See the section titled “Management — Limitations on Liability and Indemnification” appearing elsewhere in this prospectus.
Exchange Listing
Our common stock is currently not listed on any securities exchange. We have applied to have our common stock approved for listing on Nasdaq under the symbol “CTY.”
Transfer Agent and Registrar
On the completion of this offering, the transfer agent and registrar for our common stock will be Computershare Trust Company, N.A. The transfer agent’s address is 150 Royall Street, Suite 101, Canton, MA 02021.
SHARES ELIGIBLE FOR FUTURE SALE
Prior to this offering, there has been no public market for our common stock. Future sales of substantial amounts of our common stock, including shares issued on the exercise of outstanding options, in the public market after this offering, or the possibility of these sales or issuances occurring, could adversely affect the prevailing market price for our common stock or impair our ability to raise equity capital. Although we have applied to list our common stock on Nasdaq, we cannot assure you that there will be an active public market for our common stock.
Following the completion of this offering, based on our shares outstanding as of June 30, 2026, a total of 50,332,279 shares of common stock (which includes 1,964,421 shares of restricted common stock subject to repurchase or forfeiture) will be outstanding, assuming no exercise of the underwriters’ option to purchase additional shares and no exercise of outstanding options. Of the outstanding shares, all of the shares sold in this offering will be freely tradable.
All remaining shares of common stock held by existing stockholders immediately prior to the completion of this offering will be “restricted securities” as such term is defined in Rule 144. These restricted securities were issued and sold by us, or will be issued and sold by us, in private transactions and are eligible for public sale only if registered under the Securities Act or if they qualify for an exemption from registration under the Securities Act, including the exemptions provided by Rule 144 or Rule 701, summarized below.
Rule 144
In general, under Rule 144 as currently in effect, once we have been subject to public company reporting requirements of Section 13 or Section 15(d) of the Exchange Act for at least 90 days, an eligible stockholder is entitled to sell such shares without complying with the manner of sale, volume limitation or notice provisions of Rule 144, subject to compliance with the public information requirements of Rule 144. To be an eligible stockholder under Rule 144, such stockholder must not be deemed to have been one of our affiliates for purposes of the Securities Act at any time during the 90 days preceding a sale and must have beneficially owned the shares proposed to be sold for at least six months, including the holding period of any prior owner other than our affiliates. If such a person has beneficially owned the shares proposed to be sold for at least one year, including the holding period of any prior owner other than our affiliates, then such person is entitled to sell such shares without complying with any of the requirements of Rule 144, subject to the expiration of the lock-up agreements described below.
In general, under Rule 144, as currently in effect, our affiliates or persons selling shares on behalf of our affiliates are entitled to sell shares on expiration of the lock-up agreements described below. Beginning 90 days after the date of this prospectus, within any three-month period, such stockholders may sell a number of shares that does not exceed the greater of:
•
1% of the number of shares of common stock then outstanding, which will equal approximately 503,322 shares immediately after this offering, assuming no exercise of the underwriters’ option to purchase additional shares of common stock from us; or
•
the average weekly trading volume of our common stock on Nasdaq during the four calendar weeks preceding the filing of a notice on Form 144 with respect to such sale.
Sales under Rule 144 by our affiliates or persons selling shares on behalf of our affiliates are also subject to certain manner of sale provisions and notice requirements and to the availability of current public information about us.
Rule 701
Rule 701 under the Securities Act (“Rule 701”) generally allows a stockholder who was issued shares under a written compensatory plan or contract and who is not deemed to have been an affiliate of our company during the immediately preceding 90 days, to sell these shares in reliance on Rule 144, but without being required to comply with the public information, holding period, volume limitation or notice provisions of Rule 144. Rule 701 also permits affiliates of our company to sell their Rule 701 shares
under Rule 144 without complying with the holding period requirements of Rule 144. All holders of Rule 701 shares, however, are required by that rule to wait until 90 days after the date of this prospectus before selling those shares under Rule 701, subject to the expiration of the lock-up agreements described below.
Form S-8 Registration Statements
We intend to file one or more registration statements on Form S-8 under the Securities Act with the SEC to register the offer and sale of shares of our common stock that are issuable under the 2023 Plan, the 2026 Plan and the ESPP. These registration statements will become effective immediately upon filing. Shares covered by these registration statements will then be eligible for sale in the public markets, subject to vesting restrictions, any applicable lock-up agreements described below, and Rule 144 limitations applicable to affiliates.
Lock-Up Arrangements
We, all of our directors and officers, and the holders of substantially all of our capital stock and securities convertible into or exchangeable for our capital stock have entered into lock-up agreements with the underwriters and/or are subject to market standoff agreements or other agreements with us, which prevents them from selling any of our common stock or any securities convertible into or exercisable or exchangeable for common stock for a period of not less than 180 days from the date of this prospectus without the prior written consent of Goldman Sachs & Co. LLC and Jefferies LLC, subject to certain exceptions. See the section titled “Underwriting” appearing elsewhere in this prospectus for more information.
Registration Rights
Upon completion of this offering, certain holders of our securities will be entitled to various rights with respect to registration of their shares under the Securities Act. Registration of these shares under the Securities Act would result in these shares becoming fully tradable without restriction under the Securities Act immediately upon the effectiveness of the registration. See the section titled “Description of Capital Stock — Registration Rights” appearing elsewhere in this prospectus for more information.
MATERIAL U.S. FEDERAL INCOME TAX CONSEQUENCES FOR NON-U.S. HOLDERS
The following discussion is a summary of material U.S. federal income tax consequences to non-U.S. holders (as defined below) of the ownership and disposition of our common stock issued pursuant to this offering. This discussion is based on the Internal Revenue Code of 1986, as amended (the “Code”), Treasury Regulations promulgated thereunder, published rulings and administrative pronouncements of the U.S. Internal Revenue Service (the “IRS”), and judicial decisions, all as in effect on the date hereof. These authorities are subject to differing interpretations and may change, possibly with retroactive effect, resulting in U.S. federal income tax consequences different from those discussed below. We have not requested a ruling from the IRS with respect to the statements made and the conclusions reached in the following summary, and there can be no assurance that the IRS or a court will agree with such statements and conclusions.
This discussion is limited to non-U.S. holders who purchase our common stock pursuant to this offering and who hold our common stock as a “capital asset” within the meaning of Section 1221 of the Code (generally, property held for investment). This discussion is not a complete analysis of all potential U.S. federal income tax consequences relating thereto, does not address the potential application of the Medicare contribution tax on net investment income, any alternative minimum tax, or the special tax accounting rules under Section 451(b) of the Code, and also does not address any U.S. federal non-income tax consequences, such as estate or gift tax consequences, or any tax consequences arising under any state, local or non-U.S. tax laws. This discussion does not address all of the U.S. federal income tax consequences that may be relevant to a non-U.S. holder in light of such non-U.S. holder’s particular circumstances. This discussion also does not consider any specific facts or circumstances that may be relevant to non-U.S. holders subject to special rules under the U.S. federal income tax laws, including:
•
certain former citizens, or long-term residents of the United States;
•
partnerships or other entities or arrangements treated as pass-through or disregarded entities for U.S. federal income tax purposes (and investors therein);
•
“controlled foreign corporations”;
•
“passive foreign investment companies”;
•
“foreign controlled foreign corporations”;
•
corporations that accumulate earnings to avoid U.S. federal income tax;
•
banks, mutual funds, financial institutions, investment funds, insurance companies, brokers, dealers or traders in stock, securities, commodities or currencies;
•
tax-exempt organizations and governmental organizations;
•
tax-qualified retirement plans;
•
persons who acquire our common stock through the exercise of an option or otherwise as compensation;
•
qualified foreign pension funds as defined in Section 897(l)(2) of the Code and entities all of the interests of which are held by qualified foreign pension funds;
•
persons that own, or have owned, actually or constructively, more than 5% of our common stock;
•
persons who have elected to mark securities to market; and
•
persons holding our common stock as part of a hedging or conversion transaction or straddle, or synthetic security or a constructive sale, or other risk reduction strategy or integrated investment.
If an entity or arrangement that is classified as a partnership for U.S. federal income tax purposes holds our common stock, the U.S. federal income tax treatment of a partner in the partnership generally will depend on the status of the partner, the activities of the partnership and certain determinations
made at the partner level. Partnerships holding our common stock and the partners in such partnerships are urged to consult their tax advisors about the particular U.S. federal income tax consequences to them of owning and disposing of our common stock.
THIS DISCUSSION IS FOR INFORMATIONAL PURPOSES ONLY AND IS NOT TAX ADVICE. PROSPECTIVE INVESTORS SHOULD CONSULT THEIR TAX ADVISORS REGARDING THE PARTICULAR U.S. FEDERAL INCOME TAX CONSEQUENCES TO THEM OF OWNING AND DISPOSING OF OUR COMMON STOCK, AS WELL AS ANY TAX CONSEQUENCES ARISING UNDER ANY STATE, LOCAL OR NON-U.S. TAX LAWS AND ANY OTHER U.S. FEDERAL TAX LAWS OR UNDER ANY APPLICABLE INCOME TAX TREATY.
Definition of Non-U.S. Holder
For purposes of this discussion, a non-U.S. holder is any beneficial owner of our common stock that is for U.S. federal income tax purposes:
•
a non-resident alien individual;
•
a corporation or any organization taxable as a corporation for U.S. federal income tax purposes that is not created or organized under the laws of the United States, any state thereof, or the District of Columbia; or
•
a foreign trust or estate, the income of which is not subject to U.S. federal income tax on a net income basis.
Distributions on Our Common Stock
As described under “Dividend Policy,” we do not currently anticipate declaring or paying, for the foreseeable future, any distributions on our capital stock. However, if we were to distribute cash or other property on our common stock, such distributions will constitute dividends for U.S. federal income tax purposes to the extent paid from our current or accumulated earnings and profits, as determined under U.S. federal income tax principles. Amounts not treated as dividends for U.S. federal income tax purposes will then constitute a return of capital and will first be applied against and reduce a holder’s tax basis in our common stock, but not below zero. Any excess will be treated as gain realized on the sale or other disposition of our common stock and will be treated as described under “— Gain on Disposition of Our Common Stock” below.
Subject to the discussions below regarding effectively connected income, backup withholding and FATCA (as defined below), dividends paid to a non-U.S. holder of our common stock generally will be subject to U.S. federal withholding tax at a rate of 30% of the gross amount of the dividends or such lower rate specified by an applicable income tax treaty. To receive the benefit of a reduced treaty rate, a non-U.S. holder must furnish to us or our withholding agent with a timely and valid IRS Form W-8BEN (in the case of individuals) or IRS Form W-8BEN-E (in the case of entities), or other appropriate form, certifying such holder’s qualification for the reduced rate. This certification must be provided to us or our withholding agent before the payment of the dividends and must be updated periodically. If the non-U.S. holder holds our common stock through a financial institution or other agent acting on the non-U.S. holder’s behalf, the non-U.S. holder will be required to provide appropriate documentation to the agent, which then will be required to provide certification to us or our withholding agent, either directly or through other intermediaries.
If a non-U.S. holder holds our common stock in connection with the conduct of a trade or business in the United States, and dividends paid on our common stock are effectively connected with such holder’s U.S. trade or business (and, if required by an applicable tax treaty, are attributable to such holder’s permanent establishment or fixed base in the United States), such non-U.S. holder generally will be exempt from U.S. federal withholding tax. To claim the exemption, the non-U.S. holder generally must furnish a valid IRS Form W-8ECI (or applicable successor form), certifying that the dividends are effectively connected with the non-U.S. Holder’s conduct of a trade or business within the United States to the applicable withholding agent.
However, any such effectively connected dividends paid on our common stock generally will be subject to U.S. federal income tax on a net income basis at the regular U.S. federal income tax rates in the same manner as if such holder were a resident of the United States. A non-U.S. holder that is a corporation also may be subject to an additional branch profits tax equal to 30% (or such lower rate specified by an applicable income tax treaty) of its effectively connected dividends, as adjusted for certain items.
Non-U.S. holders that do not provide the required certification on a timely basis, but that qualify for a reduced treaty rate, may obtain a refund or credit of any excess amounts withheld by timely filing an appropriate claim with the IRS. Non-U.S. holders should consult their tax advisors regarding any applicable income tax treaties that may provide for different rules.
Gain on Disposition of Our Common Stock
Subject to the discussions below regarding backup withholding and FATCA (as defined below), a non-U.S. holder generally will not be subject to U.S. federal income tax on any gain realized on the sale or other taxable disposition of our common stock, unless:
•
the gain is effectively connected with the non-U.S. holder’s conduct of a trade or business in the United States and, if required by an applicable income tax treaty, is attributable to a permanent establishment or fixed base maintained by the non-U.S. holder in the United States;
•
the non-U.S. holder is a nonresident alien individual who is present in the United States for a period or periods aggregating 183 days or more during the taxable year of the disposition, and certain other requirements are met; or
•
our common stock constitutes a “United States real property interest” by reason of our status as a United States real property holding corporation (“USRPHC”) for U.S. federal income tax purposes at any time within the shorter of the five-year period preceding the disposition or the non-U.S. holder’s holding period for our common stock, and our common stock is not “regularly traded” on an “established securities market” within the meaning of applicable Treasury Regulations, during the calendar year in which the sale or other disposition occurs.
Gain described in the first bullet point above generally will be subject to U.S. federal income tax on a net income basis at the regular U.S. federal income tax rates in the same manner as if such holder were a resident of the United States. A non-U.S. holder that is a corporation also may be subject to an additional branch profits tax equal to 30% (or such lower rate specified by an applicable income tax treaty) of its effectively connected earnings and profits for the taxable year, as adjusted for certain items.
Gain described in the second bullet point above will be subject to U.S. federal income tax at a flat 30% rate (or such lower rate specified by an applicable income tax treaty), but may be offset by certain U.S.-source capital losses of the non-U.S. holder (even though the individual is not considered a resident of the United States), provided that the non-U.S. holder has timely filed U.S. federal income tax returns with respect to such losses.
Determining whether we are a USRPHC depends on the fair market value of our United States real property interests relative to the fair market value of our worldwide real property interests and our other trade or business assets. We believe that we are not currently and we do not anticipate becoming a USRPHC for U.S. federal income tax purposes, although there can be no assurance we have not been, are not currently or will not in the future become a USRPHC. Even if we are treated as a USRPHC, gain realized by a non-U.S. holder on a disposition of our common stock will not be subject to U.S. federal income tax so long as (1) the non-U.S. holder owned, directly, indirectly and constructively, no more than 5% of our common stock at all times within the shorter of (a) the five-year period preceding the disposition or (b) the holder’s holding period and (2) our common stock is “regularly traded” on an “established securities market,” within the meaning of applicable U.S. Treasury Regulations. There can be no assurance that our common stock qualifies as regularly traded on an established securities market for purposes of the rules described above. Prospective investors are encouraged to consult their own tax advisors regarding the possible consequences to them if we have been, are or were to become a USRPHC.
Non-U.S. holders should consult their tax advisors regarding any applicable income tax treaties that may provide for different rules.
Information Reporting and Backup Withholding
Annual reports are required to be filed with the IRS and provided to each non-U.S. holder indicating the amount of distributions on our common stock paid to such holder and the amount of any tax withheld with respect to those distributions. These information reporting requirements apply even if no withholding was required because the distributions were effectively connected with the holder’s conduct of a U.S. trade or business, or withholding was reduced or eliminated by an applicable income tax treaty. This information also may be made available under a specific treaty or agreement with the tax authorities in the country in which the non-U.S. holder resides or is established. Backup withholding, currently at a 24% rate, generally will not apply to payments to a non-U.S. holder of distributions on or the gross proceeds of a disposition of our common stock provided the non-U.S. holder furnishes the required certification for its non-U.S. status, such as by providing a valid IRS Form W-8BEN, IRS Form W-8BEN-E or IRS Form W-8ECI, or otherwise establishes an exemption, and if the payor does not have actual knowledge, or reason to know, that the holder is a U.S. person who is not an exempt recipient.
Backup withholding is not an additional tax. If any amount is withheld under the backup withholding rules, the non-U.S. holder should consult with a U.S. tax advisor regarding the possibility of and procedure for obtaining a refund or a credit against the non-U.S. holder’s U.S. federal income tax liability, if any.
Withholding on Foreign Entities
Sections 1471 through 1474 of the Code, which are commonly referred to as FATCA, impose a U.S. federal withholding tax of 30% on certain payments made to a “foreign financial institution” (as specially defined under these rules) unless such institution enters into an agreement with the U.S. government to withhold on certain payments and to collect and provide to the U.S. tax authorities substantial information regarding certain U.S. account holders of such institution (which includes certain equity and debt holders of such institution, as well as certain account holders that are foreign entities with U.S. owners) or an exemption applies. FATCA also generally imposes a U.S. federal withholding tax of 30% on certain payments made to a “non-financial foreign entity” (as specially defined under these rules) unless such entity provides the withholding agent a certification that it does not have any “substantial United States owners” or provides information identifying certain direct and indirect U.S. owners of the entity or an exemption applies. An intergovernmental agreement between the United States and an applicable foreign country may modify these requirements. Under certain circumstances, a non-U.S. holder might be eligible for refunds or credits of such taxes. FATCA currently applies to dividends paid on our common stock and would have applied also to payments of gross proceeds from the sale or other disposition of our common stock. However, proposed regulations under FATCA provide for the elimination of the federal withholding tax of 30% applicable to gross proceeds of a sale or other disposition of property of a type that can produce U.S.-source dividends or interest. Under these proposed Treasury Regulations (which may be relied upon by taxpayers prior to finalization), FATCA withholding does not apply to gross proceeds from sales or other dispositions of our common stock.
Prospective investors are encouraged to consult with their tax advisors regarding the possible implications of FATCA on their investment in our common stock.
EACH PROSPECTIVE INVESTOR SHOULD CONSULT ITS TAX ADVISOR REGARDING THE TAX CONSEQUENCES OF ACQUIRING, OWNING AND DISPOSING OF OUR COMMON STOCK, INCLUDING THE CONSEQUENCES OF ANY RECENT AND PROPOSED CHANGE IN APPLICABLE LAW, AS WELL AS TAX CONSEQUENCES ARISING UNDER ANY STATE, LOCAL, NON-U.S. OR U.S. FEDERAL NON-INCOME TAX LAWS.
UNDERWRITING
We and the underwriters named below have entered into an underwriting agreement with respect to the shares being offered. Subject to certain conditions, each underwriter has severally agreed to purchase the number of shares indicated in the following table. Goldman Sachs & Co. LLC and Jefferies LLC are the representatives of the underwriters.
|
Underwriters
|
|
|
Number of
Shares
|
|
|
Goldman Sachs & Co. LLC
|
|
|
|
|
|
|
|
|
Jefferies LLC
|
|
|
|
|
|
|
|
|
Stifel, Nicolaus & Company, Incorporated
|
|
|
|
|
|
|
|
|
Oppenheimer & Co. Inc.
|
|
|
|
|
|
|
|
|
Total
|
|
|
|
|
9,722,222 |
|
|
The underwriters are committed to take and pay for all of the shares being offered, if any are taken, other than the shares covered by the option described below unless and until this option is exercised.
The underwriters have an option to buy up to an additional 1,458,333 shares of our common stock from us to cover sales by the underwriters of a greater number of shares than the total number set forth in the table above. They may exercise that option for 30 days. If any shares are purchased pursuant to this option, the underwriters will severally purchase shares in approximately the same proportion as set forth in the table above.
The following table shows the per share and total underwriting discounts and commissions to be paid to the underwriters by us. Such amounts are shown assuming both no exercise and full exercise of the underwriters’ option to purchase up to an additional 1,458,333 shares of our common stock from us.
| |
|
|
No Exercise
|
|
|
Full Exercise
|
|
|
Per Share
|
|
|
|
$ |
|
|
|
|
|
$ |
|
|
|
|
Total
|
|
|
|
$ |
|
|
|
|
|
$ |
|
|
|
Shares sold by the underwriters to the public will initially be offered at the initial public offering price set forth on the cover of this prospectus. Any shares sold by the underwriters to securities dealers may be sold at a discount of up to $ per share from the initial public offering price. After the initial offering of the shares, the representatives may change the offering price and the other selling terms. The offering of the shares by the underwriters is subject to their receipt and acceptance of the shares being offered and subject to the underwriters’ right to reject any order in whole or in part.
We and our officers, directors, and holders of substantially all of our capital stock and securities convertible into or exchangeable for our common stock have agreed or will agree with the underwriters, subject to certain exceptions, not to dispose of or hedge any of our or their common stock or securities convertible into or exchangeable for shares of common stock (collectively, “Lock-Up Securities”) during the period from the date of this prospectus continuing through the date 180 days after the date of this prospectus, except with the prior written consent of Goldman Sachs & Co. LLC and Jefferies LLC. See the section titled “Shares Eligible for Future Sale” for a discussion of certain transfer restrictions.
The restrictions described in the immediately preceding paragraph do not apply to our officers, directors and holders of substantially all of our capital stock and securities convertible into or exchangeable for our common stock with respect to:
Transfers of Lock-Up Securities (i) as one or more bona fide gifts or charitable contributions, or for bona fide estate planning purposes, (ii) upon death by will, testamentary document or intestate succession, (iii) if the lock-up party is a natural person, to any member of the lock-up party’s immediate family or to any trust for the direct or indirect benefit of such lock-up party or the immediate family of
such lock-up party or, if such lock-up party is a trust, to a trust, trustee (or co-trustee) or beneficiary of the trust or the estate of a beneficiary of such trust, (iv) to a corporation, partnership, limited liability company, investment fund or other entity (A) of which the lock-up party and the immediate family of the lock-up party are the legal and beneficial owner of all of the outstanding equity securities or similar interests or (B) controlled by, or under common control with, the undersigned or the immediate family of the undersigned, (v) to a nominee or custodian of a person or entity to whom a disposition or transfer would be permissible under clauses (i) through (iv) above, (vi) if the lock-up party is a corporation, partnership, limited liability company, trust or other business entity, (A) to another corporation, partnership, limited liability company or other business entity that is a subsidiary or an affiliate (as defined in Rule 405 under the Securities Act) of such lock-up party, or to any investment fund or other entity which fund or entity is controlled or managed by or under common control or common investment management with the lock-up party or affiliates of such lock-up party (including, for the avoidance of doubt, where the undersigned is a partnership, to its general partner or a successor partnership or fund, or any other funds managed by such partnership), or (B) as part of a disposition, transfer or distribution by the lock-up party to its stockholders, partners, members or other equityholders or to the estate of any such stockholders, partners, members or other equityholders, (vii) by operation of law, such as pursuant to a qualified domestic order, divorce settlement, divorce decree or separation agreement, (viii) to us upon death or disability of the lock-up party, or, if the lock-up party is one of our employees upon death, disability or termination of employment, in each case, of such employee, (ix) if the lock-up party is not one of our officers or directors, in connection with a sale of such lock-up party’s shares of common stock acquired (A) from the underwriters in this offering or (B) in open market transactions after the closing date of this offering, or (x) to us in connection with the vesting, settlement or exercise of restricted stock units, options, warrants or other rights or instruments to purchase or acquire shares of common stock (including, in each case, by way of “net” or “cashless” exercise), including any transfer to us for the payment of tax withholdings or remittance payments due as a result of the vesting, settlement or exercise of such restricted stock units, options, warrants or other rights or instruments, or in connection with the conversion of convertible securities, in all such cases pursuant to equity awards granted under a stock incentive plan or other equity award plan, or pursuant to the terms of convertible securities, each as described in this prospectus, provided that any securities received upon such vesting, settlement, exercise or conversion shall be subject to the terms of the lock-up agreement;
provided that (A) in the case of clauses (i), (ii), (iii), (iv), (v) and (vi) above, such transfer or distribution shall not involve a disposition for value, (B) in the case of clauses (i), (ii), (iii), (iv), (v), (vi) and (vii) above, it shall be a condition to the transfer or distribution that the donee, devisee, transferee or distributee, as the case may be, shall sign and deliver a lock up agreement containing the same restrictions set forth above, (C) in the case of clauses (ii), (iii), (iv), (v) and (vi) above, no filing by any party (including, without limitation, any donor, donee, devisee, transferor, transferee, distributor or distributee) under the Exchange Act or other public filing, report or announcement reporting a reduction in beneficial ownership of Lock-Up Securities shall be required or shall be voluntarily made in connection with such transfer or distribution (other than a filing on a Form 5 or a filing required pursuant to Section 13 of the Exchange Act and the rules and regulations promulgated thereunder, in each case, made after the expiration of the lock-up period), and (D) in the case of clauses (i), (vii), (viii), (ix) and (x) above, no filing under the Exchange Act or other public filing, report or announcement shall be voluntarily made, and if any such filing, report or announcement shall be legally required during the lock-up period, such filing, report or announcement shall clearly indicate in the footnotes thereto (A) the circumstances of such transfer or distribution and (B) in the case of a transfer or distribution pursuant to clauses (i) or (vii) above, that the donee, devisee, transferee or distributee has agreed to be bound by a lock-up agreement containing the same restrictions set forth above.
In addition, the lock-up party may (a) enter into a written plan meeting the requirements of Rule 10b5-1 under the Exchange Act relating to the transfer, sale or other disposition of the lock-up party’s Lock-Up Securities, if then permitted by us, provided that none of the securities subject to such plan may be transferred, sold or otherwise disposed of until after the expiration of the lock-up period and no public announcement, report or filing under the Exchange Act, or any other public filing, report or announcement, shall be voluntarily made regarding the establishment of such plan during the lock-up period, and if any such filing, report or announcement shall be legally required during the lock-up period, such filing, report or announcement shall clearly indicate in the footnotes thereto that none of the
securities subject to such plan may be transferred, sold or otherwise disposed of pursuant to such plan until after the expiration of the lock-up period, (b) transfer the lock-up party’s Lock-Up Securities to us pursuant to an agreement under which we have the option to repurchase shares or a right of first refusal with respect to transfer of such shares, provided that no filing under the Exchange Act or other public filing, report or announcement shall be voluntarily made, and if any such filing, report or announcement shall be legally required during the lock-up period, such filing, report or announcement shall clearly indicate in the footnotes thereto the circumstances of such transfer or distribution, (c) (i) transfer its Lock-Up Securities pursuant to a bona fide third-party tender offer, merger, consolidation or other similar transaction that is approved by our board of directors and made to all holders of our capital stock involving a change of control of us, in one transaction or a series of related transactions, to a person or group of affiliated persons, of shares of capital stock if, after such transfer, such person or group of affiliated persons would hold at least a majority of our outstanding voting securities (or the voting securities of the surviving entity) and (ii) enter into any lock-up, voting or similar agreement pursuant to which the undersigned may agree to transfer, sell, tender or otherwise dispose of common stock or other such securities in connection with a transaction described in clause (i); provided that in the event that such tender offer, merger, consolidation or other similar transaction is not completed, the lock-up party’s Lock-Up Securities shall remain subject to the provisions of the lock-up agreement, or (d) convert outstanding shares of our convertible preferred stock, warrants to acquire convertible preferred stock or convertible securities into shares of common stock or warrants to acquire shares of common stock, provided that any such shares received upon such conversion shall remain subject to the provisions of the lock-up agreement.
The restrictions on transfers or other dispositions by us described above do not apply to us with respect to: (i) the sale of shares of common stock pursuant to this offering, (ii) the issuance of shares of common stock or any securities (including without limitation options, warrants, restricted stock or restricted stock units) convertible into, or exercisable for, shares of common stock pursuant to any employee stock option plan, incentive plan, stock plan, dividend reinvestment plan or otherwise in equity compensation arrangements in place as of the date of the underwriting agreement and described in this prospectus, (iii) the grant of awards pursuant to employee equity-based compensation plans, incentive plans, stock plans, or other arrangements in place as of the date of the underwriting agreement and described in this prospectus, (iv) the filing of a registration statement on Form S-8 in connection with the registration of shares of common stock issuable under any employee equity-based compensation plan, incentive plan, stock plan, dividend reinvestment plan adopted and approved by our board of directors prior to the date of the underwriting agreement and described in this prospectus, and (v) the issuance of up to 5% of the outstanding shares of our common stock in connection with the acquisition of the assets of, or a majority or controlling portion of the equity of, or a joint venture with another entity in connection with its acquisition us of such entity; provided that each recipient of any shares of common stock issued or sold pursuant to (v) above enter into a lock-up agreement with the underwriters with the same restrictions set forth above.
Prior to the offering, there has been no public market for the shares of our common stock. The initial public offering price has been negotiated among us and the representatives. Among the factors to be considered in determining the initial public offering price of the shares, in addition to prevailing market conditions, will be our historical performance, estimates of the business potential and earnings prospects of us, an assessment of our management and the consideration of the above factors in relation to market valuation of companies in related businesses.
We have applied to list our common stock on the Nasdaq Global Market under the symbol “CTY.”
In connection with the offering, the underwriters may purchase and sell shares of our common stock in the open market. These transactions may include short sales, stabilizing transactions and purchases to cover positions created by short sales. Short sales involve the sale by the underwriters of a greater number of shares than they are required to purchase in the offering, and a short position represents the amount of such sales that have not been covered by subsequent purchases. A “covered short position” is a short position that is not greater than the amount of additional shares for which the underwriters’ option described above may be exercised. The underwriters may cover any covered short position by either exercising their option to purchase additional shares or purchasing shares in the open market. In
determining the source of shares to cover the covered short position, the underwriters will consider, among other things, the price of shares available for purchase in the open market as compared to the price at which they may purchase additional shares pursuant to the option described above. “Naked” short sales are any short sales that create a short position greater than the amount of additional shares for which the option described above may be exercised. The underwriters must cover any such naked short position by purchasing shares in the open market. A naked short position is more likely to be created if the underwriters are concerned that there may be downward pressure on the price of the common stock in the open market after pricing that could adversely affect investors who purchase in the offering. Stabilizing transactions consist of various bids for or purchases of common stock made by the underwriters in the open market prior to the completion of the offering.
The underwriters may also impose a penalty bid. This occurs when a particular underwriter repays to the underwriters a portion of the underwriting discount received by it because the representatives have repurchased shares sold by or for the account of such underwriter in stabilizing or short covering transactions.
Purchases to cover a short position and stabilizing transactions, as well as other purchases by the underwriters for their own accounts, may have the effect of preventing or retarding a decline in the market price of our common stock, and together with the imposition of the penalty bid, may stabilize, maintain or otherwise affect the market price of our common stock. As a result, the price of our common stock may be higher than the price that otherwise might exist in the open market. The underwriters are not required to engage in these activities and may end any of these activities at any time. These transactions may be effected on the Nasdaq Global Market, in the over-the-counter market or otherwise.
We estimate that our share of the total expenses of the offering, excluding underwriting discounts and commissions, will be approximately $4.5 million.
We have agreed to indemnify the several underwriters against certain liabilities, including liabilities under the Securities Act. The underwriters and their respective affiliates are full service financial institutions engaged in various activities, which may include sales and trading, commercial and investment banking, advisory, investment management, investment research, principal investment, hedging, market making, brokerage and other financial and non-financial activities and services. Certain of the underwriters and their respective affiliates may in the future provide a variety of these services to us and to persons and entities with relationships with us, for which they will receive customary fees and expenses. In the ordinary course of their various business activities, the underwriters and their respective affiliates, officers, directors and employees may purchase, sell or hold a broad array of investments and actively trade securities, derivatives, loans, commodities, currencies, credit default swaps and other financial instruments for their own account and for the accounts of their customers, and such investment and trading activities may involve or relate to assets, securities and/or instruments of ours (directly, as collateral securing other obligations or otherwise) and/or persons and entities with relationships with us. The underwriters and their respective affiliates may also communicate independent investment recommendations, market color or trading ideas and/or publish or express independent research views in respect of such assets, securities or instruments and may at any time hold, or recommend to clients that they should acquire, long and/or short positions in such assets, securities and instruments.
Selling Restrictions
Other than in the United States, no action has been taken by us or the underwriters that would permit a public offering of the securities offered by this prospectus in any jurisdiction where action for that purpose is required. The securities offered by this prospectus may not be offered or sold, directly or indirectly, nor may this prospectus or any other offering material or advertisements in connection with the offer and sale of any such securities be distributed or published in any jurisdiction, except under circumstances that will result in compliance with the applicable rules and regulations of that jurisdiction. Persons into whose possession this prospectus comes are advised to inform themselves about and to observe any restrictions relating to the offering and the distribution of this prospectus. This prospectus does not constitute an offer to sell or a solicitation of an offer to buy any securities offered by this prospectus in any jurisdiction in which such an offer or a solicitation is unlawful.
European Economic Area
In relation to each Member State of the European Economic Area (each, a “Relevant Member State”), an offer to the public of any securities may not be made in that Relevant Member State prior to the publication of a prospectus in relation to the securities which has been approved by the competent authority in that Relevant Member State or, where appropriate, approved in another Relevant Member State and notified to the competent authority in that Relevant Member State, all in accordance with the Prospectus Regulation, except that an offer to the public in that Relevant Member State of any securities may be made at any time under the following exemptions under the Prospectus Regulation:
(a)
to any legal entity which is a “qualified investor” as defined under the Prospectus Regulation;
(b)
to fewer than 150 natural or legal persons (other than “qualified investors” as defined under the Prospectus Regulation), subject to obtaining the prior consent of the underwriters for any such offer; or
(c)
in any other circumstances falling within Article 1(4) of the Prospectus Regulation,
provided that no such offer of securities shall result in a requirement for the company or any of the underwriters to publish a prospectus pursuant to Article 3 of the Prospectus Regulation or a supplemental prospectus pursuant to Article 23 of the Prospectus Regulation and each person who initially acquires any securities or to whom any offer is made will be deemed to have represented, warranted and agreed to and with each of the underwriters and the company that it is a qualified investor within the meaning of Article 2 of the Prospectus Regulation.
In the case of any securities being offered to a financial intermediary as that term is used in Article 1(4) of the Prospectus Regulation, each financial intermediary will also be deemed to have represented, warranted and agreed that the securities acquired by it in the offer have not been acquired on a non-discretionary basis on behalf of, nor have they been acquired with a view to their offer or resale to, persons in circumstances which may give rise to an offer of any securities to the public, other than their offer or resale in a Relevant Member State to qualified investors as so defined or in circumstances in which the prior consent of the underwriters has been obtained to each such proposed offer or resale.
For the purposes of this provision, the expression an “offer to the public” in relation to any securities in any Relevant Member State means the communication in any form and by any means of sufficient information on the terms of the offer and any securities to be offered so as to enable an investor to decide to purchase or subscribe for any securities, and the expression “Prospectus Regulation” means Regulation (EU) 2017/1129.
United Kingdom
This prospectus has been prepared on the basis that the offering of the securities falls within one of the exceptions specified in Part 1 of Schedule 1 of the Public Offers and Admissions to Trading Regulations 2024 (the “POATRs”) and, accordingly, there will not be a prospectus prepared or published for the purposes of the POATRs. This prospectus does not constitute a prospectus for the purposes of the POATRs.
An offer to the public of any shares may not be made in the United Kingdom, except that an offer to the public in the United Kingdom of any shares may be made at any time under the following exemptions:
(a)
at any time to any legal entity which is a qualified investor as defined in paragraph 15 of Schedule 1 to the POATRs;
(b)
at any time to fewer than 150 persons (other than qualified investors as defined in paragraph 15 of Schedule 1 to the POATRs) in the United Kingdom subject to obtaining the prior consent of the relevant underwriters nominated by us for any such offer; or
(c)
at any time in any other circumstances falling within Part 1 of Schedule 1 to the POATRs.
For the purposes of this provision, the expression an “offer to the public” in relation to any securities in the United Kingdom means the communication in any form and by any means of sufficient information on the terms of the offer and the securities to be offered so as to enable an investor to decide to purchase or subscribe for the securities.
Canada
The securities may be sold in Canada only to purchasers purchasing, or deemed to be purchasing, as principal that are accredited investors, as defined in National Instrument 45-106 Prospectus Exemptions or subsection 73.3(1) of the Securities Act (Ontario), and are permitted clients, as defined in National Instrument 31-103 Registration Requirements, Exemptions, and Ongoing Registrant Obligations. Any resale of the securities must be made in accordance with an exemption from, or in a transaction not subject to, the prospectus requirements of applicable securities laws.
Securities legislation in certain provinces or territories of Canada may provide a purchaser with remedies for rescission or damages if this prospectus (including any amendment thereto) contains a misrepresentation, provided that the remedies for rescission or damages are exercised by the purchaser within the time limit prescribed by the securities legislation of the purchaser’s province or territory. The purchaser should refer to any applicable provisions of the securities legislation of the purchaser’s province or territory of these rights or consult with a legal advisor.
Pursuant to section 3A.3 of National Instrument 33-105 Underwriting Conflicts (NI 33-105), the underwriters are not required to comply with the disclosure requirements of NI 33-105 regarding underwriter conflicts of interest in connection with this offering.
Hong Kong
The securities have not been offered or sold and will not be offered or sold in Hong Kong, by means of any document, other than (a) to “professional investors” as defined in the Securities and Futures Ordinance (Cap. 571 of the laws of Hong Kong) (the “SFO”) and any rules made thereunder; or (b) in other circumstances which do not result in this prospectus being a “prospectus” as defined in the Companies (Winding Up and Miscellaneous Provisions) Ordinance (Cap. 32 of the Laws of Hong Kong) (the “CO”) or which do not constitute an offer to the public within the meaning of the CO. No advertisement, invitation or document relating to the securities has been or may be issued or has been or may be in the possession of any person for the purposes of issue, whether in Hong Kong or elsewhere, which is directed at, or the contents of which are likely to be accessed or read by, the public of Hong Kong (except if permitted to do so under the securities laws of Hong Kong) other than with respect to the securities which are or are intended to be disposed of only to persons outside Hong Kong or only to “professional investors” as defined in the SFO and any rules made thereunder.
Singapore
This prospectus has not been registered as a prospectus with the Monetary Authority of Singapore. Accordingly, the securities may not be offered or sold, or made the subject of an invitation for subscription or purchase, nor may this prospectus or any other document or material in connection with the offer or sale, or invitation for subscription or purchase of the securities be circulated, whether directly or indirectly, to any person in Singapore other than (i) to an institutional investor (as defined in Section 4A of the Securities and Futures Act 2001 of Singapore, as modified or amended from time to time (the “SFA”)) pursuant to Section 274 of the SFA or (ii) to an accredited investor (as defined in Section 4A of the SFA) pursuant to and in accordance with the conditions specified in Section 275 of the SFA.
Japan
The securities have not been and will not be registered pursuant to Article 4, Paragraph 1 of the Financial Instruments and Exchange Act. Accordingly, none of the securities nor any interest therein may be offered or sold, directly or indirectly, in Japan or to, or for the benefit of, any “resident” of, Japan (which term as used herein means any person resident in Japan, including any corporation or other entity organized under the laws of Japan), or to others for re-offering or resale, directly or indirectly, in
Japan or to or for the benefit of a resident of Japan, except pursuant to an exemption from the registration requirements of, and otherwise in compliance with, the Financial Instruments and Exchange Act and any other applicable laws, regulations and ministerial guidelines of Japan in effect at the relevant time.
Australia
No placement document, prospectus, product disclosure statement or other disclosure document has been lodged with the Australian Securities and Investments Commission in relation to this offering. This prospectus does not constitute a prospectus, product disclosure statement, or other disclosure document under Chapter 6D.2 of the Corporations Act 2001 (the “Corporations Act”), and does not purport to include the information required for a prospectus, product disclosure statement or other disclosure document under the Corporations Act.
Any offer in Australia of the securities may only be made to persons (the “Exempt Investors”) who are “sophisticated investors” (within the meaning of section 708(8) of the Corporations Act), “professional investors” (within the meaning of section 708(11) of the Corporations Act) or otherwise pursuant to one or more exemptions contained in section 708 of the Corporations Act so that it is lawful to offer the securities without disclosure to investors under Chapter 6D of the Corporations Act.
The securities applied for by Exempt Investors in Australia must not be offered for sale in Australia in the period of 12 months after the date of allotment under the offering, except in circumstances where disclosure to investors under Chapter 6D of the Corporations Act would not be required pursuant to an exemption under section 708 of the Corporations Act or otherwise, or where the offer is pursuant to a disclosure document which complies with Chapter 6D of the Corporations Act. Any person acquiring shares must observe such Australian on-sale restrictions.
This prospectus contains general information only and does not take account of the investment objectives, financial situation or particular needs of any particular person. It does not contain any securities recommendations or financial product advice. Before making an investment decision, investors need to consider whether the information in this prospectus is appropriate to their needs, objectives and circumstances and, if necessary, seek expert advice on those matters.
Dubai International Financial Centre
This prospectus relates to an “Exempt Offer” in accordance with the Offered Securities Rules of the Dubai Financial Services Authority (the “DFSA”). This prospectus is intended for distribution only to persons of a type specified in the Offered Securities Rules of the DFSA. It must not be delivered to, or relied on by, any other person. The DFSA has no responsibility for reviewing or verifying any documents in connection with Exempt Offers. The DFSA has not approved this prospectus nor taken steps to verify the information set forth herein and has no responsibility for the prospectus. The securities to which this prospectus relates may be illiquid and/or subject to restrictions on their resale. Prospective purchasers of the securities should conduct their own due diligence on the securities. If you do not understand the contents of this prospectus, you should consult an authorized financial advisor.
United Arab Emirates
The securities have not been, and are not being, publicly offered, sold, promoted or advertised in the United Arab Emirates (including the Dubai International Financial Centre) other than in compliance with the laws of the United Arab Emirates (and the Dubai International Financial Centre) governing the issue, offering and sale of securities. Further, this prospectus does not constitute a public offer of securities in the United Arab Emirates (including the Dubai International Financial Centre) and is not intended to be a public offer. This prospectus has not been approved by or filed with the Central Bank of the United Arab Emirates, the Securities and Commodities Authority, Financial Services Regulatory Authority (FSRA) or the Dubai Financial Services Authority.
Switzerland
The securities may not be publicly offered in Switzerland and will not be listed on the SIX Swiss Exchange (“SIX”) or on any other stock exchange or regulated trading facility in Switzerland. This
prospectus has been prepared without regard to the disclosure standards for issuance prospectuses under art. 652a or art. 1156 of the Swiss Code of Obligations or the disclosure standards for listing prospectuses under art. 27 ff. of the SIX Listing Rules or the listing rules of any other stock exchange or regulated trading facility in Switzerland. Neither this prospectus nor any other offering or marketing material relating to the shares or the offering may be publicly distributed or otherwise made publicly available in Switzerland.
Neither this prospectus nor any other offering or marketing material relating to the offering, us or the securities has been or will be filed with or approved by any Swiss regulatory authority. In particular, this prospectus will not be filed with, and the offer of securities will not be supervised by, FINMA, and the offer of securities has not been and will not be authorized under CISA. The investor protection afforded to acquirers of interests in collective investment schemes under the CISA does not extend to acquirers of the securities.
Brazil
The offer and sale of the securities have not been and will not be registered with the Brazilian Securities Commission (Comissão de Valores Mobiliários, or “CVM”) and, therefore, will not be carried out by any means that would constitute a public offering in Brazil under CVM Resolution No. 160, dated 13 July 2022, as amended, or unauthorized distribution under Brazilian laws and regulations. The securities may only be offered to Brazilian Professional Investors (as defined by applicable CVM regulation), who may only acquire the securities through a non-Brazilian account, with settlement outside Brazil in non-Brazilian currency. The trading of these securities on regulated securities markets in Brazil is prohibited.
LEGAL MATTERS
The validity of the shares of our common stock being offered in this prospectus will be passed upon for us by Goodwin Procter LLP. Cooley LLP is representing the underwriters in this offering.
EXPERTS
The consolidated financial statements as of December 31, 2025 and 2024 and for the years then ended included in this prospectus have been so included in reliance on the report of PricewaterhouseCoopers LLP, an independent registered public accounting firm, given on the authority of said firm as experts in auditing and accounting.
WHERE YOU CAN FIND ADDITIONAL INFORMATION
We have filed with the Securities and Exchange Commission (the “SEC”) a registration statement on Form S-1 (File Number 333-299117) under the Securities Act with respect to the shares of common stock offered by this prospectus. This prospectus, which constitutes a part of the registration statement, does not contain all the information set forth in the registration statement, some of which is contained in exhibits to the registration statement as permitted by the rules and regulations of the SEC. For further information with respect to us and our common stock, we refer you to the registration statement, including the exhibits filed as a part of the registration statement. Statements contained in this prospectus concerning the contents of any contract or any other document are not necessarily complete. If a contract or document has been filed as an exhibit to the registration statement, please see the copy of the contract or document that has been filed. Each statement in this prospectus relating to a contract or document filed as an exhibit is qualified in all respects by the filed exhibit. The SEC also maintains an internet website that contains reports and other information about issuers, like us, that file electronically with the SEC. The address of that website is www.sec.gov.
We currently do not file periodic reports with the SEC. On the completion of this offering, we will be subject to the information reporting requirements of the Exchange Act, and we will file reports, proxy statements and other information with the SEC. These reports, proxy statements and other information will be available for review at the website of the SEC referred to above.
We also maintain a website at www.citytx.com. Information contained in, or accessible through, our website is not a part of this prospectus, and the inclusion of our website address in this prospectus is only as an inactive textual reference. Upon completion of this offering, you may access, free of charge, our annual reports on Form 10-K, quarterly reports on Form 10-Q, current reports on Form 8-K and amendments to those reported filed or furnished pursuant to Section 13(a) or 15(d) of the Exchange Act as soon as reasonably practicable after such material is electronically filed with, or furnished to, the SEC.
CITY THERAPEUTICS, INC.
INDEX TO THE CONSOLIDATED FINANCIAL STATEMENTS
REPORT OF INDEPENDENT REGISTERED PUBLIC ACCOUNTING FIRM
To the Board of Directors and Stockholders of City Therapeutics, Inc.
Opinion on the Financial Statements
We have audited the accompanying consolidated balance sheets of City Therapeutics, Inc. and its subsidiary (the “Company”) as of December 31, 2025 and 2024, and the related consolidated statements of operations and comprehensive loss, convertible preferred stock and stockholders’ deficit, and cash flows for the years then ended, including the related notes (collectively referred to as the “consolidated financial statements”). In our opinion, the consolidated financial statements present fairly, in all material respects, the financial position of the Company as of December 31, 2025 and 2024, and the results of its operations and its cash flows for the years then ended in conformity with accounting principles generally accepted in the United States of America.
Basis for Opinion
These consolidated financial statements are the responsibility of the Company’s management. Our responsibility is to express an opinion on the Company’s consolidated financial statements based on our audits. We are a public accounting firm registered with the Public Company Accounting Oversight Board (United States) (PCAOB) and are required to be independent with respect to the Company in accordance with the U.S. federal securities laws and the applicable rules and regulations of the Securities and Exchange Commission and the PCAOB.
We conducted our audits of these consolidated financial statements in accordance with the standards of the PCAOB. Those standards require that we plan and perform the audits to obtain reasonable assurance about whether the consolidated financial statements are free of material misstatement, whether due to error or fraud.
Our audits included performing procedures to assess the risks of material misstatement of the consolidated financial statements, whether due to error or fraud, and performing procedures that respond to those risks. Such procedures included examining, on a test basis, evidence regarding the amounts and disclosures in the consolidated financial statements. Our audits also included evaluating the accounting principles used and significant estimates made by management, as well as evaluating the overall presentation of the consolidated financial statements. We believe that our audits provide a reasonable basis for our opinion.
/s/ PricewaterhouseCoopers LLP
Boston, Massachusetts
July 14, 2026, except for the effects of the stock split discussed in Note 18 to the consolidated financial statements, as to which the date is October 9, 2026
We have served as the Company’s auditor since 2025.
CITY THERAPEUTICS, INC.
CONSOLIDATED BALANCE SHEETS
(IN THOUSANDS, EXCEPT SHARE AND PER SHARE AMOUNTS)
| |
|
|
December 31,
|
|
| |
|
|
2024
|
|
|
2025
|
|
|
Assets
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
Current assets
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
Cash and cash equivalents
|
|
|
|
$ |
126,899 |
|
|
|
|
$ |
112,793 |
|
|
|
Accounts receivable
|
|
|
|
|
1,352 |
|
|
|
|
|
1,209 |
|
|
|
Prepaid expenses and other current assets
|
|
|
|
|
651 |
|
|
|
|
|
2,871 |
|
|
|
Total current assets
|
|
|
|
|
128,902 |
|
|
|
|
|
116,873 |
|
|
|
Property and equipment, net
|
|
|
|
|
4,362 |
|
|
|
|
|
5,489 |
|
|
|
Operating lease right-of-use asset
|
|
|
|
|
10,516 |
|
|
|
|
|
24,857 |
|
|
|
Restricted cash
|
|
|
|
|
1,528 |
|
|
|
|
|
2,504 |
|
|
|
Other non-current assets
|
|
|
|
|
45 |
|
|
|
|
|
45 |
|
|
|
Total assets
|
|
|
|
$ |
145,353 |
|
|
|
|
$ |
149,768 |
|
|
|
Liabilities, convertible preferred stock and stockholders’ deficit
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
Current liabilities
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
Accounts payable
|
|
|
|
$ |
2,339 |
|
|
|
|
$ |
2,695 |
|
|
|
Convertible notes, current
|
|
|
|
|
10,276 |
|
|
|
|
|
10,805 |
|
|
|
Operating lease liability, current
|
|
|
|
|
3,092 |
|
|
|
|
|
5,075 |
|
|
|
Deferred revenue, current
|
|
|
|
|
3,361 |
|
|
|
|
|
11,255 |
|
|
|
Accrued interest
|
|
|
|
|
385 |
|
|
|
|
|
2,642 |
|
|
|
Accrued expenses and other current liabilities
|
|
|
|
|
1,289 |
|
|
|
|
|
5,570 |
|
|
|
Total current liabilities
|
|
|
|
|
20,742 |
|
|
|
|
|
38,042 |
|
|
|
Operating lease liability, non-current
|
|
|
|
|
8,927 |
|
|
|
|
|
20,341 |
|
|
|
Convertible notes, non-current, net
|
|
|
|
|
— |
|
|
|
|
|
31,063 |
|
|
|
Other liabilities, non-current
|
|
|
|
|
— |
|
|
|
|
|
463 |
|
|
|
Deferred revenue, non-current
|
|
|
|
|
595 |
|
|
|
|
|
— |
|
|
|
Total liabilities
|
|
|
|
|
30,264 |
|
|
|
|
|
89,909 |
|
|
|
Commitments and contingencies (Note 8)
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
Series Seed convertible preferred stock, $0.0001 par value; 4,544,990 and 4,544,990 shares authorized as of December 31, 2024 and 2025; 4,544,990 and 4,544,990 shares issued and outstanding as of December 31, 2024 and 2025; aggregate liquidation preference of $4,545 and $4,545 as of December 31, 2024 and 2025
|
|
|
|
|
4,475 |
|
|
|
|
|
4,475 |
|
|
|
Series A convertible preferred stock, $0.0001 par value; 14,295,436 and
14,295,436 shares authorized as of December 31, 2024 and 2025; 13,996,318
and 13,996,318 shares issued and outstanding as of December 31, 2024 and
2025; aggregate liquidation preference of $140,376 and $140,376 as of
December 31, 2024 and 2025
|
|
|
|
|
140,039 |
|
|
|
|
|
140,039 |
|
|
|
Stockholders’ deficit:
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
Common stock, $0.0001 par value; 30,000,000 and 30,000,000 shares
authorized as of December 31, 2024 and 2025; 6,497,239 and 7,772,189
shares issued as of December 31, 2024 and 2025; 4,476,828 and 5,690,551
shares outstanding as of December 31, 2024 and 2025
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
Additional paid-in capital
|
|
|
|
|
1,174 |
|
|
|
|
|
4,449 |
|
|
|
Accumulated deficit
|
|
|
|
|
(30,599) |
|
|
|
|
|
(89,104) |
|
|
|
Total stockholders’ deficit
|
|
|
|
|
(29,425) |
|
|
|
|
|
(84,655) |
|
|
|
Total liabilities, convertible preferred stock and stockholders’ deficit
|
|
|
|
$ |
145,353 |
|
|
|
|
$ |
149,768 |
|
|
The accompanying notes are an integral part of these consolidated financial statements
CITY THERAPEUTICS, INC.
CONSOLIDATED STATEMENTS OF OPERATIONS AND COMPREHENSIVE LOSS
(IN THOUSANDS, EXCEPT SHARE AND PER SHARE AMOUNTS)
| |
|
|
Years ended December 31,
|
|
| |
|
|
2024
|
|
|
2025
|
|
|
Collaboration revenue
|
|
|
|
$ |
2,778 |
|
|
|
|
$ |
13,562 |
|
|
|
Operating expenses:
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
Research and development(1)
|
|
|
|
|
25,682 |
|
|
|
|
|
58,420 |
|
|
|
General and administrative(2)
|
|
|
|
|
6,186 |
|
|
|
|
|
15,064 |
|
|
|
Total operating expenses
|
|
|
|
|
31,868 |
|
|
|
|
|
73,484 |
|
|
|
Loss from operations
|
|
|
|
|
(29,090) |
|
|
|
|
|
(59,922) |
|
|
|
Other income:
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
Interest income
|
|
|
|
|
2,397 |
|
|
|
|
|
4,208 |
|
|
|
Interest expense
|
|
|
|
|
(448) |
|
|
|
|
|
(2,257) |
|
|
|
Other expense, net
|
|
|
|
|
(695) |
|
|
|
|
|
(534) |
|
|
|
Total other income, net
|
|
|
|
|
1,254 |
|
|
|
|
|
1,417 |
|
|
|
Net loss and comprehensive loss
|
|
|
|
$ |
(27,836) |
|
|
|
|
$ |
(58,505) |
|
|
|
Net loss per share, basic and diluted
|
|
|
|
$ |
(7.45) |
|
|
|
|
$ |
(11.34) |
|
|
|
Weighted-average common shares outstanding, basic and diluted
|
|
|
|
|
3,738,335 |
|
|
|
|
$ |
5,157,781 |
|
|
(1)
Includes related party amounts of $3.6 million and $5.3 million for research and development expenses for the years ended December 31, 2024 and 2025, respectively. See Note 17 — Related Parties.
(2)
Includes related party amounts of $0.7 million and $1.1 million for general and administrative expenses for the years ended December 31, 2024 and 2025, respectively. See Note 17 — Related Parties.
The accompanying notes are an integral part of these consolidated financial statements
CITY THERAPEUTICS, INC.
CONSOLIDATED STATEMENTS OF CONVERTIBLE PREFERRED STOCK AND STOCKHOLDERS’ DEFICIT
(IN THOUSANDS, EXCEPT SHARE AMOUNTS)
| |
|
|
Series Seed
Convertible
Preferred
Stock
|
|
|
Series A
Convertible
Preferred
Stock
|
|
|
|
Common Stock
|
|
|
Additional
Paid-In
Capital
|
|
|
Accumulated
Deficit
|
|
|
Total
Stockholders’
Deficit
|
|
| |
|
|
Shares
|
|
|
Amount
|
|
|
Shares
|
|
|
Amount
|
|
|
|
Shares
|
|
|
Amount
|
|
|
Balance as of January 1, 2024
|
|
|
|
|
4,519,990 |
|
|
|
|
$ |
4,451 |
|
|
|
|
|
— |
|
|
|
|
$ |
— |
|
|
|
|
|
|
3,341,093 |
|
|
|
|
$ |
— |
|
|
|
|
$ |
45 |
|
|
|
|
$ |
(2,763) |
|
|
|
|
$ |
(2,718) |
|
|
|
Issuance of Series Seed convertible
preferred stock, net of issuance costs of
$1
|
|
|
|
|
25,000 |
|
|
|
|
|
24 |
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
Issuance of Series A convertible preferred
stock, net of issuance costs of $336
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
|
|
13,435,354 |
|
|
|
|
|
134,414 |
|
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
Conversion of convertible debt to Series A
convertible preferred stock
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
|
|
560,964 |
|
|
|
|
|
5,625 |
|
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
Issuance of common stock in exchange for settlement of antidilution provision
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
|
|
|
32,424 |
|
|
|
|
|
— |
|
|
|
|
|
124 |
|
|
|
|
|
— |
|
|
|
|
|
124 |
|
|
|
Vesting of restricted common stock
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
|
|
|
1,103,311 |
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
Stock-based compensation expense
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
|
|
1,005 |
|
|
|
|
|
— |
|
|
|
|
|
1,005 |
|
|
|
Net loss
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
|
|
(27,836) |
|
|
|
|
|
(27,836) |
|
|
|
Balance as of December 31, 2024
|
|
|
|
|
4,544,990 |
|
|
|
|
$ |
4,475 |
|
|
|
|
|
13,996,318 |
|
|
|
|
$ |
140,039 |
|
|
|
|
|
|
4,476,828 |
|
|
|
|
$ |
— |
|
|
|
|
$ |
1,174 |
|
|
|
|
$ |
(30,599) |
|
|
|
|
$ |
(29,425) |
|
|
|
Issuance of common stock from exercised
options
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
|
|
|
421,841 |
|
|
|
|
|
— |
|
|
|
|
|
278 |
|
|
|
|
|
— |
|
|
|
|
|
278 |
|
|
|
Vesting of restricted common stock
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
|
|
|
791,882 |
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
Stock-based compensation expense
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
|
|
2,997 |
|
|
|
|
|
— |
|
|
|
|
|
2,997 |
|
|
|
Net loss
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
|
|
(58,505) |
|
|
|
|
|
(58,505) |
|
|
|
Balance as of December 31, 2025
|
|
|
|
|
4,544,990 |
|
|
|
|
$ |
4,475 |
|
|
|
|
|
13,996,318 |
|
|
|
|
$ |
140,039 |
|
|
|
|
|
|
5,690,551 |
|
|
|
|
$ |
— |
|
|
|
|
$ |
4,449 |
|
|
|
|
$ |
(89,104) |
|
|
|
|
$ |
(84,655) |
|
|
The accompanying notes are an integral part of these consolidated financial statements
CITY THERAPEUTICS, INC.
CONSOLIDATED STATEMENTS OF CASH FLOWS
(IN THOUSANDS)
| |
|
|
Years ended December 31,
|
|
| |
|
|
2024
|
|
|
2025
|
|
|
Cash flows from operating activities
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
Net loss
|
|
|
|
$ |
(27,836) |
|
|
|
|
$ |
(58,505) |
|
|
|
Adjustments to reconcile net loss to net cash used in operating activities
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
Depreciation
|
|
|
|
|
424 |
|
|
|
|
|
1,172 |
|
|
|
Stock-based compensation
|
|
|
|
|
1,005 |
|
|
|
|
|
2,997 |
|
|
|
Non-cash interest expense
|
|
|
|
|
64 |
|
|
|
|
|
2,257 |
|
|
|
Non-cash operating lease expense
|
|
|
|
|
626 |
|
|
|
|
|
314 |
|
|
|
Gain on lease modification
|
|
|
|
|
— |
|
|
|
|
|
(1,024) |
|
|
|
Change in fair value of convertible notes
|
|
|
|
|
866 |
|
|
|
|
|
1,606 |
|
|
|
Convertible note discount amortization
|
|
|
|
|
(27) |
|
|
|
|
|
21 |
|
|
|
Gain on sale of property and equipment
|
|
|
|
|
(131) |
|
|
|
|
|
(26) |
|
|
|
Changes in operating assets and liabilities:
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
Accounts receivable
|
|
|
|
|
(1,352) |
|
|
|
|
|
143 |
|
|
|
Prepaid expenses and other current assets
|
|
|
|
|
(624) |
|
|
|
|
|
(2,222) |
|
|
|
Accounts payable, accrued expenses and other current liabilities
|
|
|
|
|
2,537 |
|
|
|
|
|
4,591 |
|
|
|
Deferred revenue
|
|
|
|
|
3,956 |
|
|
|
|
|
7,299 |
|
|
|
Operating lease liabilities
|
|
|
|
|
101 |
|
|
|
|
|
(234) |
|
|
|
Net cash used in operating activities
|
|
|
|
|
(20,391) |
|
|
|
|
|
(41,611) |
|
|
|
Cash flows from investing activities
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
Purchases of property and equipment
|
|
|
|
|
(3,537) |
|
|
|
|
|
(2,405) |
|
|
|
Proceeds from sale of property and equipment
|
|
|
|
|
158 |
|
|
|
|
|
27 |
|
|
|
Net cash used in investing activities
|
|
|
|
|
(3,379) |
|
|
|
|
|
(2,378) |
|
|
|
Cash flows from financing activities
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
Proceeds from issuance of convertible notes payable
|
|
|
|
|
15,000 |
|
|
|
|
|
30,000 |
|
|
|
Payment of convertible notes payable issuance costs
|
|
|
|
|
— |
|
|
|
|
|
(35) |
|
|
|
Proceeds from exercise of stock options
|
|
|
|
|
— |
|
|
|
|
|
894 |
|
|
|
Proceeds from issuance of Series Seed convertible preferred stock
|
|
|
|
|
25 |
|
|
|
|
|
— |
|
|
|
Payment of Series Seed convertible preferred stock issuance costs
|
|
|
|
|
(1) |
|
|
|
|
|
— |
|
|
|
Proceeds from issuance of Series A convertible preferred stock
|
|
|
|
|
134,750 |
|
|
|
|
|
— |
|
|
|
Payment of Series A convertible preferred stock issuance costs
|
|
|
|
|
(336) |
|
|
|
|
|
— |
|
|
|
Net cash provided by financing activities
|
|
|
|
|
149,438 |
|
|
|
|
|
30,859 |
|
|
|
Net increase (decrease) in cash, cash equivalents, and restricted cash
|
|
|
|
|
125,668 |
|
|
|
|
|
(13,130) |
|
|
|
Cash, cash equivalents, and restricted cash at beginning of the period
|
|
|
|
|
2,759 |
|
|
|
|
|
128,427 |
|
|
|
Cash, cash equivalents, and restricted cash at end of the period
|
|
|
|
$ |
128,427 |
|
|
|
|
$ |
115,297 |
|
|
|
Supplemental disclosure of noncash investing and financing activities:
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
Property and equipment included in accounts payable and accrued expenses
|
|
|
|
|
211 |
|
|
|
|
|
107 |
|
|
|
Conversion of convertible debt into Series A convertible preferred stock
|
|
|
|
|
5,625 |
|
|
|
|
|
— |
|
|
|
Acquisition of leasehold improvements
|
|
|
|
|
1,066 |
|
|
|
|
|
— |
|
|
|
Reclassification from early exercise liability, non-current to early exercise liability, current
|
|
|
|
|
— |
|
|
|
|
|
379 |
|
|
|
Settlement of antidilution obligation
|
|
|
|
|
124 |
|
|
|
|
|
— |
|
|
|
Operating lease right-of-use assets obtained in exchange for operating
lease liabilities
|
|
|
|
|
— |
|
|
|
|
|
25,055 |
|
|
The accompanying notes are an integral part of these consolidated financial statements
CITY THERAPEUTICS, INC.
NOTES TO CONSOLIDATED FINANCIAL STATEMENTS
1. Nature of Business
Organization
City Therapeutics, Inc. (the Company or City) is a clinical-stage biotechnology company developing the next generation of RNAi therapeutics designed to silence the expression of disease-relevant proteins, with the aim of delivering meaningful and durable benefits for patients in need.
The Company was incorporated in the state of Delaware on July 17, 2023, and has a principal office in Cambridge, Massachusetts.
Liquidity and Capital Resources
Since inception, the Company has devoted substantially all of its efforts to research and development activities, advancing development of the portfolio of programs and platforms, raising capital, developing and maintaining its intellectual property, hiring personnel, and providing general and administrative support for these operations. The Company is subject to risks and uncertainties common to companies in the biotechnology industry, including but not limited to, risks of failure of preclinical studies and clinical trials, development and manufacturing of potential future product candidates, obtaining regulatory approval for potential future product candidates, competition from substitute products, the need to successfully commercialize and gain market acceptance of its potential future product candidates, dependence on key personnel, protection of proprietary technology, compliance with government regulations, development by competitors of technological innovations and the ability to secure additional capital to fund operations.
There can be no assurance that the Company’s research and development efforts will be successfully completed, that adequate protection for the Company’s intellectual property will be obtained, that any products developed will obtain necessary government regulatory approval or that any approved products will be commercially viable. Even if the Company’s product development efforts are successful, it is uncertain when, if ever, the Company will generate significant revenue from product sales. The Company operates in an environment of rapid change in technology and substantial competition from pharmaceutical and biotechnology companies. In addition, the Company is dependent upon the services of its employees and consultants.
The Company has incurred net losses of $27.8 million and $58.5 million during the years ended December 31, 2024 and 2025, respectively. As of December 31, 2024 and 2025, the Company had an accumulated deficit of $30.6 million and $89.1 million, respectively. As of December 31, 2025, the Company had cash, cash equivalents, and restricted cash of $115.3 million. To date, the Company’s financing has primarily been through the sale of its Series Seed, Series A and Series B convertible preferred stock, as well as capital received from its collaboration arrangements with Bausch + Lomb Ireland Limited (Bausch + Lomb) and Biogen MA Inc. (Biogen).
The Company has evaluated whether there are conditions and events, considered in the aggregate, that raise substantial doubt about the Company’s ability to continue as a going concern within one year after the date that these consolidated financial statements are issued. As of the issuance date of the consolidated financial statements, the Company expects its cash and cash equivalents, together with proceeds received from the Series B convertible preferred stock, will be sufficient to fund its operating expenses and capital expenditure requirements for at least the next twelve months from the date these consolidated financial statements were available to be issued. The future viability of the Company beyond that point is dependent on its ability to raise additional capital to finance its operations.
Until such time that the Company can generate significant product revenues, if ever, the Company expects to seek additional funding through private and public equity financings, additional collaborations, strategic alliances and marketing, distribution or licensing arrangements. The Company is seeking to complete an initial public offering (IPO) of its common stock. Upon the completion of a qualifying public
CITY THERAPEUTICS, INC.
NOTES TO CONSOLIDATED FINANCIAL STATEMENTS (Continued)
1. Nature of Business (Continued)
offering on specified terms, the Company’s outstanding convertible preferred stock will automatically convert into shares of common stock (see Note 10). In the event the Company does not complete an IPO, the Company may not be able to obtain financing on acceptable terms, or at all, and the Company may not be able to enter into additional collaborations or other arrangements. The terms of any financing may adversely affect the holdings or the rights of the Company’s stockholders.
If the Company is unable to obtain sufficient funding, the Company will be forced to delay, scale back or discontinue some or all of its research and development programs, product portfolio expansion efforts or commercialization efforts, which could adversely affect its business prospects. Although management continues to pursue these plans, there is no assurance that the Company will be successful in obtaining sufficient funding on terms acceptable to the Company to fund continuing operations, if at all.
2. Basis of Presentation and Summary of Significant Accounting Policies
Basis of Presentation
The accompanying consolidated financial statements have been prepared in conformity with accounting principles generally accepted in the United States of America (GAAP). Any reference in these notes to applicable guidance is meant to refer to the authoritative GAAP as found in the Accounting Standards Codification (ASC) and Accounting Standards Update (ASU) of the Financial Accounting Standards Board (FASB).
Principles of Consolidation
The accompanying consolidated financial statements include the accounts of City Therapeutics, Inc. and its wholly owned subsidiary, City Therapeutics Securities Corporation. All intercompany balances and transactions have been eliminated in consolidation.
Segment Data
The Company views its operations and manages its business as a single segment for the purposes of assessing performance and making operating decisions. The Company’s chief operating decision maker (CODM) reviews the Company’s financial information on a consolidated basis for purposes of assessing performance and allocating resources. The Company’s CODM is the Chief Executive Officer (CEO).
Use of Estimates
The preparation of the Company’s consolidated financial statements in conformity with GAAP requires management to make estimates and assumptions that affect the reported amounts of assets and liabilities, the disclosure of contingent liabilities at the date of the consolidated financial statements, and the reported amounts of expenses during the reporting period. Significant estimates and assumptions reflected in these consolidated financial statements include, but are not limited to, estimates and assumptions for the valuation of common stock and stock-based compensation expense, the incremental borrowing rate related to the Company’s lease, fair value measurement of convertible notes, and the accrual of research and development expenses. The Company bases its estimates on historical experience, known trends and other market-specific or other relevant factors that it believes to be reasonable under the circumstances. On an ongoing basis, management evaluates its estimates when there are changes in circumstances, facts and experience. Changes in estimates are recorded in the period in which they become known. Actual results could differ from those estimates, and any such differences may be material to the Company’s consolidated financial statements.
CITY THERAPEUTICS, INC.
NOTES TO CONSOLIDATED FINANCIAL STATEMENTS (Continued)
2. Basis of Presentation and Summary of Significant Accounting Policies (Continued)
Cash, Cash Equivalents, and Restricted Cash
Cash represents funds in the Company’s operating and savings bank accounts. The Company considers all highly liquid investments with an original maturity of three months or less when purchased to be cash equivalents.
As of December 31, 2024 and 2025, restricted cash consists of cash placed in a restricted bank account as required under the terms of the Company’s lease agreement, which is recorded as restricted cash within the Company’s consolidated balance sheets.
The following table provides a reconciliation of cash, cash equivalents, and restricted cash as reported within the Company’s consolidated balance sheets to cash, cash equivalents, and restricted cash in the consolidated statements of cash flows (in thousands):
| |
|
|
December 31,
|
|
| |
|
|
2024
|
|
|
2025
|
|
|
Cash and cash equivalents
|
|
|
|
$ |
126,899 |
|
|
|
|
$ |
112,793 |
|
|
|
Restricted cash
|
|
|
|
|
1,528 |
|
|
|
|
|
2,504 |
|
|
|
Total cash, cash equivalents, and restricted cash
|
|
|
|
$ |
128,427 |
|
|
|
|
$ |
115,297 |
|
|
Concentration of Credit Risk
The Company has no significant off-balance sheet concentrations of credit risk, such as foreign currency exchange contracts, option contracts, or other hedging arrangements. Financial instruments that potentially subject the Company to concentrations of credit risk primarily consist of cash and cash equivalents balances in several accounts with two financial institutions which often exceed federally insured limits. The Company places its cash in accredited financial institutions and the Company does not believe that it is subject to credit risk beyond the normal credit risk associated with commercial banking relationships.
Fair Value Measurements
The accounting guidance defines fair value, establishes a consistent framework for measuring fair value and expands disclosure for each major asset and liability category measured at fair value on either a recurring or non-recurring basis. Fair value is defined as an exit price, representing the amount that would be received to sell an asset or paid to transfer a liability in an orderly transaction between market participants. As such, fair value is a market-based measurement that should be determined based on assumptions that market participants would use in pricing an asset or liability. As a basis for considering such assumptions, the accounting guidance establishes a three-tier fair value hierarchy, which prioritizes the inputs used in measuring fair value as follows:
Level 1: Observable inputs such as quoted prices in active markets.
Level 2: Inputs, other than the quoted prices in active markets that are observable either directly or indirectly.
Level 3: Unobservable inputs in which there is little or no market data, which require the reporting entity to develop its own assumptions.
As permitted under ASC 825, Financial Instruments (ASC 825), the Company has elected the fair value option to account for its convertible notes issued. The Company believes that measuring the notes at fair value better reflects the economics of the instruments because the notes contain multiple embedded
CITY THERAPEUTICS, INC.
NOTES TO CONSOLIDATED FINANCIAL STATEMENTS (Continued)
2. Basis of Presentation and Summary of Significant Accounting Policies (Continued)
features, including conversion rights, that would otherwise require separate accounting analyses. The fair value option simplifies the accounting for the instruments and provides users of the financial statements with information that is more representative of the current value of the Company’s obligations. In accordance with ASC 825, the Company records the convertible notes at fair value at each reporting period until settlement, with changes in fair value recorded in the consolidated statements of operations and comprehensive loss. As a result of applying the fair value option, direct costs and fees related to the convertible notes were recognized in earnings as incurred and not deferred.
The Company was obligated to issue to the Ohio State Innovation Foundation (OSIF) shares of the Company’s common stock pursuant to an antidilution provision (Antidilution Obligation) included within the Company’s exclusive license agreement with OSIF (Note 9). At the time of the agreement with OSIF, the Company recorded the Antidilution Obligation at fair value and remeasured the liability at each reporting period, with changes in fair value recognized in the consolidated statement of operations and comprehensive loss. The OSIF Antidilution Obligation was settled in April 2024.
The Company’s convertible notes are classified within the Level 3 designation and are recorded at fair value on a recurring basis until settled. The carrying amounts of the Company’s financial instruments, including cash and cash equivalents, prepaid expenses and other current assets, accounts payable, and accrued liabilities, approximate fair value due to their relatively short maturities.
Accounts Receivable and Allowance for Credit Losses
Accounts receivable are stated at the historical carrying amount net of write-offs and allowance for credit losses in accordance with ASC 326, Financial Instruments — Credit Losses. The Company makes judgments as to its ability to collect outstanding receivables and provides an estimate of expected credit losses for receivables when collection becomes doubtful. The Company’s receivables relate to amounts reimbursed under its research and license agreements. The Company believes that credit risk associated with its research and license partners is not significant. The Company analyzes the actual payment history of its customer, the aging of receivables, current customer-specific developments and economic trends to estimate the reserve for current expected credit losses. To date, the Company has not had any write-offs of accounts receivable, and the Company did not have an allowance for expected credit losses as of December 31, 2024 and 2025.
Deferred Offering Costs
The Company capitalizes certain legal, professional, accounting, and other third-party fees that are directly associated with in-process equity financings as deferred offering costs until such equity financings are consummated. After consummation of the equity financing, these costs are recorded as a reduction in the proceeds from the offering, either as a reduction of the carrying amount of the convertible preferred stock or in stockholders’ deficit as a reduction of additional paid-in capital generated as a result of the offering. Should the in-process equity financing be abandoned, the deferred offering costs are expensed immediately as a charge to operating expenses in the consolidated statement of operations and comprehensive loss. As of December 31, 2024 and 2025, no deferred offering costs were recorded within prepaid expenses and other current assets and other non-current assets within the consolidated balance sheets.
CITY THERAPEUTICS, INC.
NOTES TO CONSOLIDATED FINANCIAL STATEMENTS (Continued)
2. Basis of Presentation and Summary of Significant Accounting Policies (Continued)
Property and Equipment, Net
Property and equipment consists of laboratory equipment, computers, furniture and office equipment, and is recorded at cost, less accumulated depreciation. Depreciation is calculated using the straight-line method over the estimated useful lives of the assets as follows:
|
Asset
|
|
|
Estimated Useful Life
|
|
|
Lab equipment
|
|
|
5 years
|
|
|
Computer equipment and software
|
|
|
3 years
|
|
|
Furniture and office equipment
|
|
|
7 years
|
|
|
Leasehold improvements
|
|
|
Shorter of useful life or remaining lease term
|
|
Maintenance and repairs that do not improve or extend the lives of the respective assets are expensed to operations as incurred, while costs of major additions and betterments are capitalized. The Company capitalizes laboratory equipment used for research and development if it has alternative future use in research and development or otherwise. Upon retirement or sale, the cost of assets disposed and the related accumulated depreciation and amortization are removed from the accounts and any resulting gain or loss is included in loss from operations.
Impairment of Long-Lived Assets
Long-lived assets consist of property and equipment and operating lease right-of-use assets. The Company continually evaluates long-lived assets for potential impairment whenever events or changes in circumstances indicate that the carrying value of the assets may not be fully recoverable. Factors that the Company considers in deciding when to perform an impairment review include significant underperformance of the business in relation to expectations, significant negative industry or economic trends and significant changes or planned changes in the use of the assets. If an impairment review is performed to evaluate a long-lived asset group for recoverability, the Company compares the carrying values of the asset group to the expected future undiscounted cash flows that the asset group is expected to generate from the use and eventual disposition of the asset group. An impairment loss would be recognized when estimated undiscounted future cash flows expected to result from the use of an asset group are less than its carrying amount of that asset group. The amount of the impairment loss to be recognized would be calculated by the excess of the carrying value of the impaired asset group over the asset group’s fair value. The Company did not recognize any impairment losses on long-lived assets during the years ended December 31, 2024 and 2025.
Leases
The Company has an operating lease for office and lab space. At the inception of a lease arrangement, the Company determines whether the arrangement is or contains a lease based on specific facts and circumstances present, the existence of an identified asset(s), if any, and control over the use of the identified asset(s), if applicable. Operating lease liabilities and their corresponding right-of-use assets are recorded based on the present value of future lease payments over the expected lease term. The expected lease term includes noncancelable lease periods and, when applicable, periods covered by an option to extend the lease if the Company is reasonably certain to exercise that option, as well as periods covered by an option to early terminate the lease if the Company is reasonably certain not to exercise that option. The interest rate implicit in lease contracts is typically not readily determinable. As such, the Company uses an incremental borrowing rate, which is the rate incurred to borrow, on a collateralized basis over a similar term, an amount equal to the lease payments in a similar economic environment.
CITY THERAPEUTICS, INC.
NOTES TO CONSOLIDATED FINANCIAL STATEMENTS (Continued)
2. Basis of Presentation and Summary of Significant Accounting Policies (Continued)
The Company does not recognize assets or liabilities for leases with lease terms of less than 12 months. Lease payments for short-term leases are recorded to operating expense on a straight-line basis over the lease term and variable lease payments are recorded in the period in which the obligation for those payments is incurred.
The Company has elected, as a policy, to combine lease and non-lease components as a single component for all leases. Operating leases are recognized on the accompanying consolidated balance sheets as operating lease right-of-use assets, operating lease liabilities, current, and operating lease liabilities, non-current. Fixed rents are included in the calculation of the lease balances while variable costs paid for certain operating and passthrough costs are excluded and reflected as expense in the period they are paid. Lease expense is recognized over the expected term on a straight-line basis.
Commitments and Contingencies
The Company may have certain contingent liabilities that arise in the ordinary course of business. The Company accrues a liability for such matters when it is probable that future expenditures will be made and can be reasonably estimated. If some amount within a range of loss appears at the time to be a better estimate than any other amount within the range, the Company accrues that amount. When no amount within the range is a better estimate than any other amount the Company accrues the minimum amount in the range.
Stock-based Compensation
The Company accounts for its stock-based compensation awards in accordance with ASC Topic 718, Compensation — Stock Compensation (ASC 718), which requires all stock-based payments to be recognized in the consolidated statements of operations and comprehensive loss based on their fair values. Under the Company’s 2023 Stock Plan, as amended (2023 Plan), the Company can grant stock-based awards to employees, directors, and non-employee consultants in the form of stock options to purchase shares of its common stock or restricted stock awards. Compensation expense is measured at estimated fair value on the grant date, and is included as compensation expense over the requisite service period.
The fair value of each stock option award is estimated on the date of grant using the Black-Scholes option pricing model. As there is no public market for its common stock, the Company determined the volatility for awards granted based on an analysis of reported data for a group of guideline public companies. The expected volatility is based on the historical volatility of this group of guideline public companies. The Company expects to continue to do so until such time as it has adequate historical data regarding the volatility of its own traded stock price. The expected term of the Company’s stock options has been determined utilizing the “simplified” method for awards that qualify as “plain-vanilla” options. The risk-free interest rate is determined by reference to the U.S. Treasury yield curve in effect at the time of grant of the award for time periods approximately equal to the expected term of the award. The Company has not paid, and does not anticipate paying, cash dividends on its common stock; therefore, the expected dividend yield is assumed to be zero.
As there is no public market for the Company’s common stock, the Company determined the fair value of the shares of its common stock underlying its share-based awards by considering a number of objective and subjective factors, including third-party valuations of the Company’s common stock, the valuation of comparable companies, the Company’s operating and financial performance, and general and industry-specific economic outlook, amongst other factors. The assumptions underlying these valuations represented management’s best estimate, with the assistance of a third-party valuation specialist, which involved inherent uncertainties and the application of management’s judgment. These third-party valuations were performed in accordance with the guidance outlined in the American
CITY THERAPEUTICS, INC.
NOTES TO CONSOLIDATED FINANCIAL STATEMENTS (Continued)
2. Basis of Presentation and Summary of Significant Accounting Policies (Continued)
Institute of Certified Public Accountants’ Accounting and Valuation Guide, Valuation of Privately Held Company Equity Securities Issued as Compensation. As a result, if the Company had used different assumptions or estimates, the fair value of the Company’s common stock and its stock-based compensation expense could have been materially different.
Compensation expense for awards granted to employees and directors with service-based vesting conditions is recognized on a straight-line basis over the requisite service period of the respective award, which is generally the vesting period of the award. Compensation expense for awards granted to non-employees is recognized in the same period and manner as if the Company had paid cash for the services provided, which is generally the vesting period of the award. Compensation for stock options with performance-based vesting conditions is recognized when it is probable that the performance condition will be achieved. The Company accounts for forfeitures of stock-based awards as they occur.
The Company’s stock option plan allows for early exercise of stock options prior to vesting. Shares of common stock issued upon early exercise of unvested stock options are considered legally outstanding but are not deemed to be issued for accounting purposes until the underlying options vest. The consideration received for early-exercised shares is recorded as an early exercise liability on the consolidated balance sheets and reclassified to additional paid-in capital as the underlying options vest in accordance with the original vesting schedule. The Company can repurchase the shares at the original exercise price and in the event the employee leaves prior to vesting, any unvested shares are subject to repurchase by the Company at the lower of the exercise price or the fair market value of the restricted shares at the time the repurchase right is exercised.
The Company classifies stock-based compensation expense in its consolidated statements of operations and comprehensive loss in the same manner in which the award recipient’s salary and related costs are classified or in which the award recipient’s service payments are classified.
Convertible Preferred Stock
The Company records convertible preferred stock at fair value on the dates of issuance, net of issuance costs. Upon the occurrence of certain events that are outside the Company’s control, including a deemed liquidation event, such as a merger or consolidation, or a sale or transfer of substantially all of the Company’s assets, holders of the convertible preferred stock can cause redemption for cash.
Therefore, the convertible preferred stock is classified outside of permanent stockholders’ deficit as mezzanine equity (temporary equity) and is presented in the mezzanine section of the consolidated balance sheets as events triggering the redemption for cash are not solely within the Company’s control. The carrying values of the convertible preferred stock are adjusted to their liquidation preferences if and when it becomes probable that such a liquidation event will occur. No accretion was recognized for the years ended December 31, 2024 and 2025 as the contingent events that could give rise to redemption were not deemed probable.
Comprehensive Loss
Comprehensive loss is comprised of net loss and other comprehensive income (loss). For the years ended December 31, 2024 and 2025 there were no differences between net loss and comprehensive loss.
Net loss per share
The Company follows the two-class method when computing net loss per share as the Company has issued convertible preferred stock which are participating securities. The two-class method determines net loss per share for each class of common and participating securities, according to dividends declared
CITY THERAPEUTICS, INC.
NOTES TO CONSOLIDATED FINANCIAL STATEMENTS (Continued)
2. Basis of Presentation and Summary of Significant Accounting Policies (Continued)
or accumulated and participation rights in undistributed earnings. The two-class method requires income available to common stockholders for the period to be allocated between common and participating securities based upon their respective rights.
The Company’s convertible preferred stock does not contractually require the holders of such shares to participate in losses of the Company. Accordingly, in periods in which the Company reported a net loss attributable to common stockholders, such losses are not allocated to such participating securities. In periods in which the Company reported a net loss attributable to common stockholders, diluted net loss per share attributable to common stockholders is the same as basic net loss per share attributable to common stockholders, since dilutive common stock are not assumed to have been issued if their effect is antidilutive. The Company reported a net loss attributable to common stockholders for the years ended December 31, 2024 and 2025.
Basic net loss per share attributable to common stockholders is computed by dividing net loss attributable to common stockholders by the weighted-average number of shares of common stock outstanding during the period. Diluted net loss per share attributable to common stockholders is computed by dividing the diluted net loss attributable to common stockholders by the weighted-average number of shares of common stock outstanding for the period, including potential dilutive common stock.
Acquired In-Process Research and Development Expenses
The Company measures and recognizes asset acquisitions or licenses to intellectual property that are not deemed to be business combinations based on the cost to acquire or license the asset or group of assets, which includes transaction costs. Goodwill is not recognized in asset acquisitions or transactions to license intellectual property. In an asset acquisition or license to intellectual property, the cost allocated to acquire in-process research and development (IPR&D) with no alternative future use is recognized as expense on the acquisition date.
Upfront and milestone payments made are accrued for and expensed when the achievement of the milestone is probable up to the point of regulatory approval. Milestone payments made upon regulatory approval are capitalized and amortized over the remaining useful life of the related product.
There were no acquired IPR&D expenses recognized for the years ended December 31, 2024 and 2025.
Patent Costs
The Company expenses patent and patent application costs and related legal costs for the prosecution and maintenance of such patents and patent applications, including patents and patent applications the Company in-licenses, as incurred and classifies such costs as general and administrative expenses in the accompanying consolidated statements of operations and comprehensive loss.
Revenue Recognition
The Company recognizes revenue in accordance with ASC Topic 606, Revenue from Contracts with Customers (ASC 606). The Company enters into collaboration agreements and certain other agreements that are within the scope of ASC 606, under which the Company licenses, or grants an option to license rights to certain of the Company’s product candidates and performs research and development services in connection with such arrangements. The terms of these arrangements typically include payment of one or more of the following: non-refundable, up-front fees; reimbursement of research and development costs; development, regulatory and commercial sales milestone payments; and royalties on net sales of licensed products.
CITY THERAPEUTICS, INC.
NOTES TO CONSOLIDATED FINANCIAL STATEMENTS (Continued)
2. Basis of Presentation and Summary of Significant Accounting Policies (Continued)
Under ASC 606, an entity recognizes revenue when its customer obtains control of promised goods or services, in an amount that reflects the consideration which the entity expects to receive in exchange for those goods or services. To determine the appropriate amount of revenue to be recognized for arrangements determined to be within the scope of ASC 606, the Company performs the following five steps: (i) identification of the customer and contract with the customer; (ii) identification of promised goods or services and which are performance obligations; (iii) measurement of the transaction price, including the constraint on variable consideration; (iv) allocation of the transaction price to the performance obligations; and (v) recognition of revenue when (or as) the Company satisfies each performance obligation. The Company only applies the five-step model to contracts when it is probable that the entity will collect consideration it is entitled to in exchange for the goods or services it transfers to the customer.
The promised goods or services in the Company’s arrangements typically consist of a license, or option to license, rights to the Company’s intellectual property or research and development services. The Company provides options for additional items in such arrangements, which are accounted for as separate contracts when the customer elects to exercise such options, unless the option provides a material right to the customer. Performance obligations are promised goods or services in a contract to transfer a distinct good or service to the customer and are considered distinct when (i) the customer can benefit from the good or service on its own or together with other readily available resources and (ii) the promised good or service is separately identifiable from other promises in the contract. In assessing whether promised good or services are distinct, the Company considers factors such as the stage of development of the underlying intellectual property, the capabilities of the customer to develop the intellectual property on its own or whether the required expertise is readily available and whether the goods or services are integral or dependent to other goods or services in the contract.
The Company estimates the transaction price based on the amount expected to be received for transferring the promised goods or services in the contract. The consideration may include fixed consideration or variable consideration. At the inception of each arrangement that includes variable consideration, the Company evaluates the amount of potential payment and the likelihood that the payments will be received.
The Company utilizes either the most likely amount method or expected value method to estimate the amount expected to be received based on which method best predicts the amount expected to be received. The amount of variable consideration that is included in the transaction price may be constrained and is included in the transaction price only to the extent that it is probable that a significant reversal in the amount of the cumulative revenue recognized will not occur in a future period.
The Company’s contracts often include development and regulatory milestone payments that are assessed under the most likely amount method and constrained if it is probable that a significant revenue reversal would occur. Milestone payments that are not within the Company’s control or the licensee’s control, such as regulatory approvals, are not considered probable of being achieved until those approvals are received. At the end of each reporting period, the Company re-evaluates the probability of achievement of such development and regulatory milestones and any related constraint, and if necessary, adjusts its estimate of the overall transaction price. Any such adjustments are recorded on a cumulative catch-up basis, which would affect collaboration revenue in the period of adjustment.
For arrangements that include sales-based royalties, including milestone payments based on the level of sales, and the license is deemed to be the predominant item to which the royalties relate, the Company recognizes revenue when the related sales occur. To date, the Company has not recognized any royalty revenue resulting from any of the Company’s arrangements.
The Company allocates the transaction price based on the estimated standalone selling price. The Company must develop assumptions that require judgment to determine the stand-alone selling price for each performance obligation identified in the contract. The Company utilizes key assumptions to
CITY THERAPEUTICS, INC.
NOTES TO CONSOLIDATED FINANCIAL STATEMENTS (Continued)
2. Basis of Presentation and Summary of Significant Accounting Policies (Continued)
determine the stand-alone selling price, which may include other comparable transactions, pricing considered in negotiating the transaction and the estimated costs. Variable consideration is allocated specifically to one or more performance obligations in a contract when the terms of the variable consideration relate to the satisfaction of the performance obligation and the resulting amounts allocated are consistent with the amounts the Company would expect to receive for the satisfaction of each performance obligation.
The consideration allocated to each performance obligation is recognized as revenue when control is transferred for the related goods or services. For performance obligations which consist of licenses and other promises, the Company utilizes judgment to assess the nature of the combined performance obligation to determine whether the combined performance obligation is satisfied over time or at a point in time and, if over time, the appropriate method of measuring progress. The Company recognizes revenue based on the cost-to-cost method for revenue recognition and total revenue amounts are based on actual costs incurred relative to total costs expected to be incurred.
The Company receives payments from its customers based on billing schedules established in each contract. Up-front payments and fees are recorded as deferred revenue upon receipt or when due until the Company performs its obligations under these arrangements. Amounts are recorded as accounts receivable when the Company’s right to consideration is unconditional.
Research and Development
Research and development costs are expensed as incurred. Research and development costs include salaries, stock-based compensation and other employee benefit expenses, consultants, costs of funding research performed by third parties on the Company’s behalf, lab-related supplies, and allocated facility-related expenses.
Non-refundable advance payments for goods and services to be received in the future for use in research and development activities are recorded as prepaid expenses on the accompanying consolidated balance sheets. The prepaid amounts are expensed as the related goods are delivered or the services are performed, or when it is no longer expected that the goods will be delivered, or the services rendered.
The Company records accrued liabilities for estimated costs of research and development activities conducted by third-party service providers. The Company accrues these costs based on factors such as estimates of the work completed and in accordance with agreements established with its third-party service providers under the service agreements. The Company makes significant judgments and estimates in determining the accrued liabilities balance in each reporting period. As actual costs become known, the Company adjusts its accrued liabilities. The historical accrual estimates made by the Company have not been materially different from the actual costs.
Income Taxes
Income taxes are accounted for under the asset and liability method. Deferred tax assets and liabilities are recognized for the future tax consequences attributable to differences between the financial statement carrying amounts of existing assets and liabilities and their respective tax bases and operating loss and tax credit carryforwards. Deferred tax assets and liabilities are measured using enacted tax rates expected to apply to taxable income in the years in which those temporary differences are expected to be recovered or settled. The effect on deferred tax assets and liabilities of a change in tax rates is recognized in income in the period that includes the enactment date. The Company assesses the likelihood that its deferred tax assets will be recovered from future taxable income and, to the extent it believes, based upon the weight of available evidence, that it is more likely than not that all or a portion of deferred tax assets will not be realized, a valuation allowance is established through a charge to
CITY THERAPEUTICS, INC.
NOTES TO CONSOLIDATED FINANCIAL STATEMENTS (Continued)
2. Basis of Presentation and Summary of Significant Accounting Policies (Continued)
income tax expense. Potential for recovery of deferred tax assets is evaluated by estimating the future taxable profits expected and considering prudent and feasible tax planning strategies. The Company recognizes the effect of income tax positions only if those positions are more likely than not to be sustained. Recognized income tax positions are measured at the largest amount that is greater than 50% likely to be realized. Changes in recognition or measurement are reflected in the period in which the change in judgment occurs.
The Company records interest related to unrecognized tax benefits and penalties in income tax expenses. The Company did not recognize any interest or penalties in the periods presented.
Recently Adopted Accounting Standards
In December 2023, the FASB issued ASU No. 2023-09, Income Taxes (Topic 740): Improvements to Income Tax Disclosures (ASU 2023-09). Additionally, the amendments in the ASU 2023-09 update require entities to disclose certain information about income taxes paid, rate reconciliation, income tax disaggregation, and the removal of disclosures related to temporary differences surrounding deferred tax liabilities to enhance the transparency and decision usefulness of income tax disclosures. For public entities, the amendments in this update are effective for fiscal years beginning after December 15, 2024 and for entities other than public entities, the amendments in this update are effective for fiscal years beginning after December 15, 2025. Early adoption is permitted for any annual periods for which financial statements have not been issued or made available for issuance. The Company elected to early adopt the standard in 2025 on a retrospective basis. See Note 14 — Income Taxes, for further disclosure.
In November 2024, the FASB issued ASU No. 2024-04, Debt — Debt with Conversion and Other Options (Subtopic 470-20): Induced Conversions of Convertible Debt Instruments (ASU 2024-04), which clarifies the requirements for determining whether the settlement of a convertible debt instrument should be accounted for as an induced conversion or as a debt extinguishment. ASU 2024-04 is effective for the Company for annual reporting periods beginning after December 15, 2025, and interim reporting periods within those annual periods, with early adoption permitted for entities that have adopted the amendments in ASU 2020-06. Adoption did not have a material impact on the Company’s consolidated financial statements and related disclosures.
In September 2025, the FASB issued ASU No. 2025-07 Derivatives Scope Refinements and Scope Clarification for Share-Based Noncash Consideration from a Customer in a Revenue Contract (ASU 2025-07). The amendments provide for a new scope exception to the derivatives guidance for underlyings based on the operations or activities specific to one of the parties to the contract, and also clarifies that share-based noncash consideration received from a customer as consideration for the transfer of goods or services in a revenue contract is subject to the revenue guidance and not the financial instruments guidance unless and until the company’s right to receive or retain the share-based noncash consideration is unconditional as defined in ASU 2025-07. The standard is effective for annual reporting periods beginning after December 15, 2026, including interim reporting periods within those annual periods, with early adoption permitted. The Company adopted this new standard on a prospective basis as of January 1, 2025. Adoption did not have a material impact on the Company’s consolidated financial statements and related disclosures.
Recently Issued Accounting Standards
In November 2024, the FASB issued ASU 2024-03, Income Statement — Reporting Comprehensive Income — Expense Disaggregation Disclosures (Subtopic 220-40): Disaggregation of Income Statement Expenses (ASU 2024-03), which is intended to provide more detailed information about specified categories of expenses included in certain expense captions presented on the statement of operations
CITY THERAPEUTICS, INC.
NOTES TO CONSOLIDATED FINANCIAL STATEMENTS (Continued)
2. Basis of Presentation and Summary of Significant Accounting Policies (Continued)
and comprehensive loss. The guidance in ASU 2024-03 is effective for fiscal years beginning after December 15, 2026. Early adoption is permitted. The amendments may be applied either (1) prospectively to financial statements issued for periods after the effective date of ASU 2024-03 or (2) retrospectively to all prior periods presented in the financial statements. The Company is currently evaluating the impact that the adoption of ASU 2024-03 may have on its consolidated financial statements and related disclosures for fiscal years beginning after December 15, 2026.
3. Property and Equipment, Net
Property and equipment, net consisted of the following (in thousands):
| |
|
|
December 31,
|
|
| |
|
|
2024
|
|
|
2025
|
|
|
Computer and equipment
|
|
|
|
$ |
24 |
|
|
|
|
$ |
198 |
|
|
|
Furniture and fixtures
|
|
|
|
|
16 |
|
|
|
|
|
16 |
|
|
|
Lab equipment
|
|
|
|
|
3,680 |
|
|
|
|
|
5,805 |
|
|
|
Leasehold improvements
|
|
|
|
|
1,066 |
|
|
|
|
|
1,066 |
|
|
|
Total property and equipment, gross
|
|
|
|
|
4,786 |
|
|
|
|
|
7,085 |
|
|
|
Less: accumulated depreciation
|
|
|
|
|
(424) |
|
|
|
|
|
(1,596) |
|
|
|
Property and equipment, net
|
|
|
|
$ |
4,362 |
|
|
|
|
$ |
5,489 |
|
|
Total depreciation expense for the years ended December 31, 2024 and 2025 was $0.4 million and $1.2 million, respectively.
4. Accrued Expenses and Other Current Liabilities
Accrued expenses and other current liabilities consisted of the following (in thousands):
| |
|
|
December 31,
|
|
| |
|
|
2024
|
|
|
2025
|
|
|
Payroll and employee-related costs
|
|
|
|
$ |
887 |
|
|
|
|
$ |
2,480 |
|
|
|
Research and development costs
|
|
|
|
|
204 |
|
|
|
|
|
2,162 |
|
|
|
Other
|
|
|
|
|
198 |
|
|
|
|
|
928 |
|
|
|
Total accrued expenses and other current liabilities
|
|
|
|
$ |
1,289 |
|
|
|
|
$ |
5,570 |
|
|
5. Convertible Notes
Original Notes
In January and February 2024, the Company issued a series of four convertible notes (the Original Notes) to ARCH Venture Fund XII L.P. (ARCH) and one of the Company’s founders (the Founder) for aggregate cash proceeds of $5.0 million. The Original Notes accrued interest at a rate of 7% per annum. The Company elected the fair value option to account for the Original Notes.
In April 2024, in connection with the execution of the Company’s Series A Convertible Preferred Stock Purchase Agreement (Series A Agreement), the outstanding principal and accrued interest under the Original Notes were automatically converted into 560,964 shares of Series A convertible preferred stock at a discounted conversion price of $9.02655 per share. This price reflects a 10% discount to the
CITY THERAPEUTICS, INC.
NOTES TO CONSOLIDATED FINANCIAL STATEMENTS (Continued)
5. Convertible Notes (Continued)
$10.0295 per share purchase price paid by new investors in the Series A financing. The conversion of the Original Notes was accounted for as an extinguishment of debt. The Company recognized the fair value of the Series A convertible preferred stock issued upon conversion in temporary equity on the consolidated balance sheet. As a result, the Company recorded an adjustment of $0.6 million on the Original Notes which was included in total other income, net, in the consolidated statements of operations and comprehensive loss.
B+L Note
In July 2024, the Company, in connection with the research and license agreement entered into with Bausch + Lomb (Note 9), issued a convertible note (B+L Note) to Bausch + Lomb for cash proceeds of $10.0 million. The B+L Note was issued at fair value. The B+L Note bears interest at a rate of 8% per annum and matures on July 8, 2027, unless earlier converted into equity securities of the Company. The Company elected the fair value option to account for the B+L Note. Accordingly, the Company recorded the B+L Note at fair value upon issuance and at each reporting period until settlement, with changes in fair value recorded in the consolidated statements of operations and comprehensive loss.
The principal and accrued interest under the B+L Note will automatically convert into equity securities upon the initial closing of the Company’s next equity financing in which the Company raises at least $50.0 million of cash proceeds. The holder of the B+L Note may also elect to convert the note into the equity securities sold in the next equity financing in which the Company raises less than $50.0 million in cash proceeds. The number of shares issued upon conversion will equal the outstanding principal and accrued interest divided by a conversion price equal to 85% of the price paid per share by cash investors in such financing.
The note purchase agreement contains customary events of default, such as failure to pay amounts when due, bankruptcy, and certain material adverse changes. Upon the occurrence of an event of default (and after any applicable cure periods), all outstanding amounts under the B+L Note may be declared immediately due and payable.
For the years ended December 31, 2024 and 2025, the Company recognized $0.4 million and $0.8 million of interest expense related to the B+L Note, respectively. The Company recognized a loss on change in fair value within total other income, net, in the consolidated statements of operations and comprehensive loss for the B+L Note in the amount of $0.3 million and $0.5 million for the years ended December 31, 2024 and 2025, respectively. The fair value of the B+L Note as of December 31, 2024 and 2025 was $10.3 million and $10.8 million, respectively.
Biogen Note
In May 2025, the Company, in connection with the research collaboration and license agreement entered into with Biogen (Note 9), issued a convertible note (Biogen Note) to Biogen for cash proceeds of $30.0 million. The Biogen Note was issued at fair value. The Biogen Note bears interest at a rate of 8% per annum and matures on July 1, 2027, unless earlier converted into equity securities of the Company. The Company elected the fair value option to account for the Biogen Note. Accordingly, the Company recorded the Biogen Note at fair value upon issuance and at each reporting period until settlement, with changes in fair value recorded in the consolidated statements of operations and comprehensive loss.
The principal and accrued interest under the Biogen Note will automatically convert into shares of the Company’s equity securities upon the earlier of (i) the initial closing of the Company’s next qualified equity financing in which the Company raises at least $100.0 million of cash proceeds or (ii) the closing of the Company’s IPO. In the case of a qualified equity financing, the Biogen Note will convert into shares of preferred stock issued in such financing. In the case of an IPO, the Biogen Note will convert
CITY THERAPEUTICS, INC.
NOTES TO CONSOLIDATED FINANCIAL STATEMENTS (Continued)
5. Convertible Notes (Continued)
into shares of common stock issued in such offering. If the Biogen Note remains outstanding as of June 30, 2027, the outstanding principal and accrued interest will automatically convert into shares of preferred stock of the Company’s most recently completed equity financing. In each case, the number of shares issued upon conversion will equal the outstanding principal and accrued interest divided by a conversion price equal to 85% of the price paid per share by cash investors in the applicable financing.
The note purchase agreement contains customary events of default, such as failure to pay amounts when due, bankruptcy, and certain material adverse changes. Upon the occurrence of an event of default (and after any applicable cure periods), all outstanding amounts under the Biogen Note may be declared immediately due and payable.
For the year ended December 31, 2025, the Company recognized $1.5 million of interest expense related to the Biogen Note. The Company recognized a loss on change in fair value within total other income, net, in the consolidated statements of operations and comprehensive loss for the Biogen Note in the amount of $1.1 million for the year ended December 31, 2025. The loss on change in fair value primarily reflects the passage of time as the note accreted toward its expected conversion value, with a remaining estimated time to settlement of 1.25 years as of December 31, 2025. The fair value of the Biogen Note as of December 31, 2025 was $31.1 million.
6. Fair Value of Financial Instruments
The following tables present information about the Company’s financial assets and liabilities that are measured at fair value on a recurring basis and indicate the fair value hierarchy of the valuation techniques the Company utilized to determine such fair value (in thousands):
| |
|
|
Fair Value Measurements as of December 31, 2024:
|
|
| |
|
|
Level 1
|
|
|
Level 2
|
|
|
Level 3
|
|
|
Total
|
|
|
Liabilities
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
Convertible debt, net of discount
|
|
|
|
$ |
— |
|
|
|
|
$ |
— |
|
|
|
|
$ |
10,276 |
|
|
|
|
$ |
10,276 |
|
|
|
Total liabilities
|
|
|
|
$ |
— |
|
|
|
|
$ |
— |
|
|
|
|
$ |
10,276 |
|
|
|
|
$ |
10,276 |
|
|
| |
|
|
Fair Value Measurements as of December 31, 2025:
|
|
| |
|
|
Level 1
|
|
|
Level 2
|
|
|
Level 3
|
|
|
Total
|
|
|
Liabilities
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
Convertible debt, net of discount
|
|
|
|
$ |
— |
|
|
|
|
$ |
— |
|
|
|
|
$ |
41,868 |
|
|
|
|
$ |
41,868 |
|
|
|
Total liabilities
|
|
|
|
$ |
— |
|
|
|
|
$ |
— |
|
|
|
|
$ |
41,868 |
|
|
|
|
$ |
41,868 |
|
|
There have been no transfers between fair value levels during the years ended December 31, 2024 and 2025.
Convertible Notes
As further described in Note 5, the Company issued convertible notes to ARCH, a Company Founder and Bausch + Lomb in January, February and July 2024, respectively. The convertible note issued to Bausch + Lomb was in connection with a research collaboration and license agreement. In May 2025, the Company issued a convertible note to Biogen in connection with a research collaboration and license agreement. The Company elected the fair value option for each of the convertible note issuances. The fair value of each of the convertible notes was estimated using a scenario-based analysis that estimates the fair value based on the probability-weighted present value of expected future investment returns, considering each of the possible outcomes available to the noteholder.
CITY THERAPEUTICS, INC.
NOTES TO CONSOLIDATED FINANCIAL STATEMENTS (Continued)
6. Fair Value of Financial Instruments (Continued)
The notes issued to ARCH and the Founder, as described in Note 5, were converted into Series A convertible preferred stock in April 2024. The notes issued to Bausch + Lomb and Biogen remain outstanding as of December 31, 2025.
The Company adjusts the carrying value of its convertible notes to their estimated fair value at each reporting date, with any related increases or decreases recorded as change in fair value of convertible notes within total other income, net, in the consolidated statement of operations and comprehensive loss.
For the year ended December 31, 2024, the Company recognized a net loss on change in fair value of convertible notes of $0.9 million in the consolidated statements of operations and comprehensive loss, consisting of a loss on change in fair value of the B+L Note of $0.3 million and a loss on change in fair value of the ARCH and Founder notes of $0.6 million. For the year ended December 31, 2025, the Company recognized a net loss on change in fair value of convertible notes of $1.6 million in the consolidated statements of operations and comprehensive loss, consisting of a loss on change in fair value of the Biogen Note of $1.1 million and a loss on change in fair value of the B+L Note of $0.5 million.
The following table summarizes information about the significant unobservable inputs used in the fair value measurements for the B+L Note and the Biogen Note as of December 31, 2024 and 2025, respectively:
| |
|
|
Fair value as
of initial
valuation
date
(in thousands)
|
|
|
Fair value as
of period end
(in thousands)
|
|
|
Expected
settlement
options
(and relative
weighting)
|
|
|
Key
unobservable
inputs
|
|
|
Range
|
|
|
B+L Note
|
|
|
|
$ |
10,000 |
|
|
|
|
$ |
10,276 |
|
|
|
Equity
financing (95%)
|
|
|
Estimated
time to
settlement
Discount rate
|
|
|
0.5 years
18.17%
|
|
| |
|
|
Fair value as
of initial
valuation
date
(in thousands)
|
|
|
Fair value as
of period end
(in thousands)
|
|
|
Expected
settlement
options
(and relative
weighting)
|
|
|
Key
unobservable
inputs
|
|
|
Range
|
|
|
B+L Note
|
|
|
|
$ |
10,000 |
|
|
|
|
$ |
10,805 |
|
|
|
Equity
financing (95%)
|
|
|
Estimated
time to
settlement
Discount rate
|
|
|
0.42 years
18.17%
|
|
|
Biogen Note
|
|
|
|
$ |
30,000 |
|
|
|
|
$ |
31,063 |
|
|
|
Equity
financing (95%)
|
|
|
Estimated
time to
settlement
Discount rate
|
|
|
1.25 years
14.39%
|
|
CITY THERAPEUTICS, INC.
NOTES TO CONSOLIDATED FINANCIAL STATEMENTS (Continued)
6. Fair Value of Financial Instruments (Continued)
The following table provides a reconciliation of all liabilities measured at fair value using Level 3 significant unobservable inputs (in thousands):
| |
|
|
Convertible Notes
|
|
|
Balance at January 1, 2024
|
|
|
|
$ |
— |
|
|
|
Issuance of convertible notes on January 5, 2024
|
|
|
|
|
2,500 |
|
|
|
Issuance of convertible notes on February 15, 2024
|
|
|
|
|
2,500 |
|
|
|
Issuance of convertible note, net on July 8, 2024
|
|
|
|
|
9,973 |
|
|
|
Change in fair value of convertible notes
|
|
|
|
|
866 |
|
|
|
Non-cash conversion of convertible notes to Series A convertible preferred
stock
|
|
|
|
|
(5,563) |
|
|
|
Balance at December 31, 2024
|
|
|
|
$ |
10,276 |
|
|
|
Issuance of convertible note, net on May 23, 2025
|
|
|
|
|
29,965 |
|
|
|
Amortization of issuance costs
|
|
|
|
|
21 |
|
|
|
Change in fair value of convertible notes
|
|
|
|
|
1,606 |
|
|
|
Balance at December 31, 2025
|
|
|
|
$ |
41,868 |
|
|
During the years ended December 31, 2024 and 2025, there were no transfers between levels.
7. Leases
In December 2023, the Company entered into a lease of office and laboratory space at 399 Binney Street in Cambridge, Massachusetts, which became the Company’s corporate headquarters. In June 2025, the Company entered into the First Amendment to Lease (the Amendment), which expanded the leased premises to include the entire second floor and a portion of the fourth floor, provided for the surrender of the original first floor and penthouse mechanical premises and extended the lease term through September 30, 2031. Following the Amendment, the Company’s leased premises consist of approximately 51,726 rentable square feet. The lease is recorded in the accompanying consolidated balance sheets as an operating lease right-of-use asset and operating lease liability as of December 31, 2024 and 2025.
Under the terms of the Amendment, the lease term for the retained and substitute premises expires on September 30, 2031. The lease includes a renewal option which extends the term of the lease for an additional 36 months, however, it has been determined that it is not reasonably certain that the Company will exercise this option. The monthly rent payments under the lease, which include base rent charges of approximately $0.5 million per month, are subject to yearly rent increases of 3% through the end of the lease term. Base rent is fully abated during the months of December 2025, December 2026, and December 2027. The Company is responsible for paying a pro-rata share of costs incurred for common area maintenance, real estate taxes, and property insurance related to the leased space, which are considered variable lease payments and expensed as incurred. The total rent payments to be paid over the noncancelable term of this lease are $39.6 million.
CITY THERAPEUTICS, INC.
NOTES TO CONSOLIDATED FINANCIAL STATEMENTS (Continued)
7. Leases (Continued)
The elements of operating lease expense were as follows (in thousands):
| |
|
|
December 31,
|
|
| |
|
|
2024
|
|
|
2025
|
|
|
Lease cost
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
Operating lease cost
|
|
|
|
$ |
3,356 |
|
|
|
|
$ |
4,551 |
|
|
|
Variable lease cost
|
|
|
|
|
1,320 |
|
|
|
|
|
1,868 |
|
|
|
Total lease cost
|
|
|
|
$ |
4,676 |
|
|
|
|
$ |
6,419 |
|
|
Future lease payments under noncancelable leases that have commenced as of December 31, 2025 (in thousands):
|
Years ended December 31,
|
|
|
|
|
|
2026
|
|
|
|
$ |
5,122 |
|
|
|
2027
|
|
|
|
|
5,803 |
|
|
|
2028
|
|
|
|
|
6,520 |
|
|
|
2029
|
|
|
|
|
6,716 |
|
|
|
2030
|
|
|
|
|
6,918 |
|
|
|
2031
|
|
|
|
|
5,344 |
|
|
|
Total lease payments
|
|
|
|
|
36,423 |
|
|
|
Less: imputed interest
|
|
|
|
|
(11,007) |
|
|
|
Total operating lease liabilities at December 31, 2025
|
|
|
|
$ |
25,416 |
|
|
|
Current operating lease liabilities
|
|
|
|
$ |
5,075 |
|
|
|
Non-current operating lease liabilities
|
|
|
|
|
20,341 |
|
|
|
Total
|
|
|
|
$ |
25,416 |
|
|
As of December 31, 2024 and 2025, the Company’s operating leases had a weighted-average incremental borrowing rate of 14.0% and a weighted-average remaining lease term of 4.2 years and 5.7 years, respectively.
8. Commitments and Contingencies
Litigation
Liabilities for loss contingencies arising from claims, assessments, litigation, fines, penalties, and other sources are recorded when it is probable that a liability has been incurred and the amount can be reasonably estimated. From time to time, the Company may become involved in legal proceedings arising in the ordinary course of business. For the years ended December 31, 2024 and 2025, the Company had no matters for which a loss was considered probable or reasonably possible that would require accrual or disclosure under ASC 450, Contingencies.
Indemnification
In the ordinary course of business, the Company may provide indemnification of varying scope and terms to vendors, lessors, business partners and other parties with respect to certain matters including, but not limited to, losses arising out of breach of such agreements or from intellectual property infringement claims made by third parties. In addition, the Company has entered into, or intends to
CITY THERAPEUTICS, INC.
NOTES TO CONSOLIDATED FINANCIAL STATEMENTS (Continued)
8. Commitments and Contingencies (Continued)
enter into, indemnification agreements with all members of the board of directors (Board) that will require the Company, among other things, to indemnify them against certain liabilities that may arise by reason of their status or service as directors.
The maximum potential amount of future payments the Company could be required to make under these indemnification agreements is not determinable. To date, the Company has not incurred any material costs as a result of such indemnifications. The Company is not aware of any claims under indemnification arrangements that could have a material effect on its financial position, results of operations or cash flows, and it has not accrued any liabilities related to such obligations in its consolidated financial statements as of December 31, 2024 and 2025.
9. Significant Agreements
Ohio State Innovation Foundation License Agreement
On October 30, 2023, the Company entered into an exclusive license agreement (as amended, the OSIF License Agreement) with OSIF, pursuant to which the Company has been granted an exclusive, royalty-bearing license to certain patent rights to make, have made, use, sell, and import licensed products. As consideration, the Company issued 622,801 shares of common stock, which constituted 5% of the fully diluted capitalization of the Company as of the effective date of the agreement, valued at $0.1 million. On April 17, 2024, upon the closing of the Series A convertible preferred stock financing (Note 10), the Company issued an additional 32,424 shares, valued at $0.1 million, of common stock in accordance with certain anti-dilution provisions of the OSIF License Agreement, which allowed for OSIF to maintain its applicable pro rata ownership percentage until the Company had raised a specified amount of proceeds. The Company did not incur any material costs in connection with the issuance of the shares. The OSIF License Agreement has subsequently been amended to add additional patent rights and technology disclosures to the licensed intellectual property, and the Company reimbursed OSIF for patent prosecution costs associated with such additional patent rights.
Pursuant to the OSIF License Agreement, the Company is required to pay OSIF a total of up to $4.2 million upon achievement of certain clinical development, regulatory approval, and commercial sales milestones for each licensed product. The Company is also required to pay royalties to OSIF in the low-single digit percentage range based on net sales of any licensed products and processes, along with a specified minimum annual royalty amount for each licensed product, subject to customary reductions and offsets. In addition, the Company is responsible for certain patent costs incurred after the effective date and annual maintenance fees. The OSIF License Agreement also requires the Company to pay OSIF tiered percentages of certain non-royalty sublicense consideration, ranging from a low-twenties to mid-twenties percentage at earlier stages of development and a low-tens to mid-teens percentage at later stages of development, with the applicable percentage depending on the stage of development at which the sublicense is granted. In connection with the Biogen Research Collaboration and License Agreement (Note 9), during the year ended December 31, 2025, the Company paid OSIF $0.3 million in sublicensing fees related to the upfront payment received under the agreement with Biogen. No milestone payments or net sales-based royalties had become due to OSIF through December 31, 2025. No repurchase rights of common stock issued to OSIF existed as of December 31, 2024 or 2025.
Accounting Considerations
The Company determined that the OSIF License Agreement represented an asset acquisition of IPR&D assets with no alternative future use and recognized the aggregate acquisition cost as acquired in-process research and development expense in the consolidated statement of operations and comprehensive loss. The acquisition did not qualify as a business combination as the acquisition did
CITY THERAPEUTICS, INC.
NOTES TO CONSOLIDATED FINANCIAL STATEMENTS (Continued)
9. Significant Agreements (Continued)
not include both an input and substantive processes, including an assembled workforce, that together contribute to the ability to create outputs. No acquired in-process research and development expense was recognized in connection with the OSIF License Agreement during the years ended December 31, 2024 or 2025.
Bausch + Lomb Research Collaboration and License Agreement
On July 8, 2024, the Company entered into a research collaboration and license agreement (B+L Research Collaboration and License Agreement) with Bausch + Lomb, pursuant to which (i) the Company shall undertake a research program to discover certain licensed molecules which would enable Bausch + Lomb to develop, and subsequently commercialize, one or more licensed products in ocular disease and (ii) the Company granted Bausch + Lomb an option to obtain an exclusive, worldwide license to certain Company intellectual property to develop, manufacture and commercialize licensed molecules and licensed products in ocular diseases. Simultaneously with the Company’s entry into the B+L Research Collaboration and License Agreement, the Company issued the B+L Note to Bausch + Lomb and Bausch + Lomb transferred to the Company $10.0 million in consideration (Note 5).
Pursuant to the B+L Research Collaboration and License Agreement, the Company will perform research and deliver to Bausch + Lomb a development candidate for which Bausch + Lomb may exercise their option to license. If Bausch + Lomb exercises its option, it will engage in the development, manufacturing and commercialization of licensed molecules and licensed products from the development candidate. Under the B+L Research Collaboration and License Agreement, the Company received a non-refundable upfront payment of $5.0 million from Bausch + Lomb. Over the course of the research plan, Bausch + Lomb is required to reimburse the Company for amounts incurred for full-time employees and other related costs and materials directly associated with the research plan. Additionally, the Company is eligible to receive a certain development candidate selection fee and certain other development, regulatory, commercial and sales milestones, collectively up to an aggregate of $485.0 million, along with tiered royalties ranging in the mid-single digits to low teens on net sales. If the agreement terminates and the Company later successfully commercializes a product candidate using the licensed intellectual property, the Company will owe a low-single digit royalty payment to Bausch + Lomb on future net sales for a fixed royalty term. As of December 31, 2025, no such milestone payments or royalties had become due to the Company.
Accounting Considerations
The Company assessed the B+L Research Collaboration and License Agreement and concluded that Bausch + Lomb is a customer in accordance with ASC 606. The Company identified the following promises under the contract: (i) obligation to perform research services, (ii) joint research committee participation, (iii) an option for Bausch + Lomb to obtain a license to develop, manufacture and commercialize licensed molecules and licensed products, and (iv) an option for Bausch + Lomb to request technology and Chemistry, Manufacturing, and Controls transfer services. The Company assessed the promised goods and services to determine if they are distinct performance obligations. The Company determined that the options for additional goods and services are not priced below their respective standalone selling prices, and therefore, are not material rights or performance obligations. The Company also determined that the obligation to perform research services and joint research committee participation are highly interrelated and interdependent and therefore not distinct. Accordingly, the obligation to perform research services and joint research committee participation represent one combined performance obligation, and the entire transaction price will be allocated to that single combined performance obligation.
At contract inception, the transaction price was determined to be $13.3 million, which represents the aggregate of the fixed consideration related to the upfront non-refundable payment for the B+L Research
CITY THERAPEUTICS, INC.
NOTES TO CONSOLIDATED FINANCIAL STATEMENTS (Continued)
9. Significant Agreements (Continued)
Collaboration and License Agreement and the estimated variable consideration related to cost reimbursements for the research plan. As of December 31, 2025, the Company updated its estimate of variable consideration, resulting in an increase in the total transaction price to approximately $15.0 million. The Company recognizes the transaction price as the single performance obligation is satisfied, using an input method which is based on the actual costs incurred relative to total costs expected to be incurred. The transfer of control occurs over this period and, in management’s judgment, is the best measure of progress towards satisfying the performance obligation. Costs to perform the Company’s obligations under the B+L Research Collaboration and License Agreement are recognized as research and development expenses in the period incurred.
Biogen Research Collaboration and License Agreement
On May 23, 2025, the Company entered into a research collaboration and license agreement (Biogen Research Collaboration and License Agreement) with Biogen, pursuant to which (i) the Company and Biogen shall collaborate to undertake a research plan to discover certain RNAi triggers that inhibit the expression of a target protein and conjugates of such RNAi triggers and (ii) the Company granted Biogen an exclusive, worldwide, royalty-bearing license to certain Company intellectual property to develop, manufacture and commercialize a certain licensed products in the field in the territory. Simultaneously with the Company’s entry into the Biogen Research Collaboration and License Agreement, the Company issued the Biogen Note to Biogen and Biogen transferred to the Company $30.0 million in consideration (Note 5).
Pursuant to the Biogen Research Collaboration and License Agreement, the Company will perform research activities and deliver to Biogen data packages and technology transfers to enable Biogen to select and advance a development candidate, for which Biogen may exercise its rights under the license. If Biogen selects a development candidate, it will control the development, manufacturing and commercialization of licensed compounds and licensed products. Each party is responsible for its own research costs under the agreement. Biogen has the option of selecting one additional target during the option exercise period, subject to execution of a separate agreement and payment of a specified option exercise fee. Under the Biogen Research Collaboration and License Agreement, the Company received a $16.0 million upfront payment from Biogen. The Company is also eligible to receive an additional upfront payment contingent upon the satisfaction of certain conditions relating to the Company’s upstream third-party intellectual property agreement. Additionally, the Company is eligible to receive up to an aggregate of approximately $1.0 billion in potential development, commercial, and sales milestones, along with tiered royalties ranging from the high-single digits to low teens on net sales. As of December 31, 2025, no such milestones or royalties had become due to the Company.
Accounting Considerations
The Company assessed the Biogen Research Collaboration and License Agreement and concluded that Biogen is a customer in accordance with ASC 606. The Company identified the following promises under the contract: (i) an exclusive license to develop, manufacture and commercialize licensed compounds and licensed products, (ii) an obligation to perform research activities under the research plan, (iii) joint research committee participation, (iv) technology transfer and assistance, and (v) an option for Biogen to select one additional target. The Company assessed the promised goods and services to determine if they are distinct performance obligations. The Company determined that the option for an additional target is not priced below its standalone selling price, and therefore is not a material right or performance obligation. The Company also determined that the license, obligation to perform research activities, joint research committee participation, and technology transfer and assistance are highly interrelated and interdependent and therefore not distinct. Accordingly, these promises represent one combined performance obligation, and the transaction price will be allocated to that single combined performance obligation.
CITY THERAPEUTICS, INC.
NOTES TO CONSOLIDATED FINANCIAL STATEMENTS (Continued)
9. Significant Agreements (Continued)
At contract inception, the transaction price was determined to be $16.0 million, which represents the non-refundable upfront payment under the Biogen Research Collaboration and License Agreement. The $4.0 million additional upfront payment, research milestones, development milestones, sales milestones, and royalties are variable consideration that is fully constrained at inception and therefore excluded from the initial transaction price. During the year ended December 31, 2025, one research milestone of $2.5 million became probable and was included in the transaction price, increasing the total transaction price to $18.5 million as of December 31, 2025. The Company recognizes the transaction price as the single performance obligation is satisfied, using an input method which is based on the actual costs incurred relative to total costs expected to be incurred. The transfer of control occurs over this period and, in management’s judgment, is the best measure of progress towards satisfying the performance obligation. Costs to perform the Company’s obligations under the Biogen Research Collaboration and License Agreement are recognized as research and development expenses in the period incurred.
Amounts recognized under the B+L Research Collaboration and License Agreement and Biogen Research Collaboration and License Agreement were as follows (in thousands):
| |
|
|
December 31,
|
|
| |
|
|
2024
|
|
|
2025
|
|
|
Revenue recognized from B+L Research Collaboration and License Agreement (Ocular)
|
|
|
|
$ |
2,778 |
|
|
|
|
$ |
7,123 |
|
|
|
Revenue recognized from Biogen Research Collaboration and License Agreement
(CNS)
|
|
|
|
|
— |
|
|
|
|
|
6,439 |
|
|
|
Total Revenue
|
|
|
|
$ |
2,778 |
|
|
|
|
$ |
13,562 |
|
|
|
Amounts receivable under B+L Research Collaboration and License Agreement
|
|
|
|
$ |
1,352 |
|
|
|
|
$ |
1,209 |
|
|
|
Deferred revenue, current portion
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
Deferred revenue – Bausch + Lomb, current portion
|
|
|
|
|
3,361 |
|
|
|
|
|
1,693 |
|
|
|
Deferred revenue – Biogen, current portion
|
|
|
|
|
— |
|
|
|
|
|
9,562 |
|
|
|
Total Deferred Revenue, current portion
|
|
|
|
$ |
3,361 |
|
|
|
|
$ |
11,255 |
|
|
|
Deferred revenue, non-current portion
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
Deferred revenue – Bausch + Lomb, noncurrent portion
|
|
|
|
$ |
595 |
|
|
|
|
$ |
— |
|
|
|
Deferred revenue – Biogen, noncurrent portion
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
Total Deferred Revenue, non-current portion
|
|
|
|
$ |
595 |
|
|
|
|
$ |
— |
|
|
The accounts receivable recorded as of December 31, 2024 and 2025 related to the reimbursable research and development efforts by the Company which were invoiced as incurred.
The following table summarizes the changes in deferred revenue related to the upfront non-refundable payments received as part of the B+L Research Collaboration and License Agreement and Biogen Research Collaboration and License Agreement (in thousands):
| |
|
|
December 31,
|
|
| |
|
|
2024
|
|
|
2025
|
|
|
Beginning balance
|
|
|
|
$ |
— |
|
|
|
|
$ |
3,956 |
|
|
|
Additions
|
|
|
|
|
5,000 |
|
|
|
|
|
16,000 |
|
|
|
Deductions
|
|
|
|
|
(1,044) |
|
|
|
|
|
(8,701) |
|
|
|
Ending balance
|
|
|
|
$ |
3,956 |
|
|
|
|
$ |
11,255 |
|
|
CITY THERAPEUTICS, INC.
NOTES TO CONSOLIDATED FINANCIAL STATEMENTS (Continued)
9. Significant Agreements (Continued)
The Company recognizes the following revenues from amounts included in deferred revenue as of the beginning of the period (in thousands):
| |
|
|
December 31,
|
|
| |
|
|
2024
|
|
|
2025
|
|
|
Amounts included in deferred revenue as of the beginning of the period
|
|
|
|
$ |
— |
|
|
|
|
$ |
2,078 |
|
|
10. Convertible Preferred Stock
Series Seed Convertible Preferred Stock
Beginning in September 2023 and in multiple closings through March 2024, the Company issued shares of its Series Seed convertible preferred stock pursuant to a Series Seed Convertible Preferred Stock Purchase Agreement (Series Seed Agreement) with certain investors. Pursuant to the Series Seed Agreement, the Company issued a total of 4,544,990 shares of Series Seed convertible preferred stock at a purchase price of $1.00 per share for gross proceeds of $4.5 million across multiple closings. The Company incurred $0.1 million in issuance costs associated with the Series Seed convertible preferred stock financing.
Series A Convertible Preferred Stock
Beginning in April 2024 and closing in multiple closings through December 2024, the Company issued shares of its Series A convertible preferred stock pursuant to the Series A Agreement with certain investors. Pursuant to the Series A Agreement, the Company issued an aggregate of 13,996,318 shares of Series A convertible preferred stock. Of the 13,996,318 shares issued, 13,435,354 shares were issued at a price per share of $10.0295 for aggregate gross proceeds of $134.7 million. Additionally, the Company’s outstanding convertible notes and related accrued interest, at the time of the Series A Agreement, totaling $5.1 million, converted into 560,964 shares of Series A convertible preferred stock at a price per share of $9.02655, as further described in Note 5.
The Company’s convertible preferred stock is summarized below (in thousands, except share amounts):
| |
|
|
As of December 31, 2024
|
|
| |
|
|
Total Shares
Authorized
|
|
|
Total Shares
Issued and
Outstanding
|
|
|
Common
Shares
Issuable upon
Conversion
|
|
|
Carrying
Value
|
|
|
Liquidation
Preference
|
|
|
Series Seed convertible preferred stock
|
|
|
|
|
4,544,990 |
|
|
|
|
|
4,544,990 |
|
|
|
|
|
4,991,760 |
|
|
|
|
$ |
4,475 |
|
|
|
|
$ |
4,545 |
|
|
|
Series A convertible preferred stock
|
|
|
|
|
14,295,436 |
|
|
|
|
|
13,996,318 |
|
|
|
|
|
15,372,145 |
|
|
|
|
|
140,039 |
|
|
|
|
|
140,376 |
|
|
|
Total
|
|
|
|
|
18,840,426 |
|
|
|
|
|
18,541,308 |
|
|
|
|
|
20,363,905 |
|
|
|
|
$ |
144,514 |
|
|
|
|
$ |
144,921 |
|
|
CITY THERAPEUTICS, INC.
NOTES TO CONSOLIDATED FINANCIAL STATEMENTS (Continued)
10. Convertible Preferred Stock (Continued)
| |
|
|
As of December 31, 2025
|
|
| |
|
|
Total Shares
Authorized
|
|
|
Total Shares
Issued and
Outstanding
|
|
|
Common
Shares
Issuable upon
Conversion
|
|
|
Carrying
Value
|
|
|
Liquidation
Preference
|
|
|
Series Seed convertible preferred stock
|
|
|
|
|
4,544,990 |
|
|
|
|
|
4,544,990 |
|
|
|
|
|
4,991,760 |
|
|
|
|
$ |
4,475 |
|
|
|
|
$ |
4,545 |
|
|
|
Series A convertible preferred stock
|
|
|
|
|
14,295,436 |
|
|
|
|
|
13,996,318 |
|
|
|
|
|
15,372,145 |
|
|
|
|
|
140,039 |
|
|
|
|
|
140,376 |
|
|
|
Total
|
|
|
|
|
18,840,426 |
|
|
|
|
|
18,541,308 |
|
|
|
|
|
20,363,905 |
|
|
|
|
$ |
144,514 |
|
|
|
|
$ |
144,921 |
|
|
The holders of the Series A and Series Seed convertible preferred stock have the following rights, preferences and privileges:
Conversion
All series of convertible preferred stock are convertible at any time at the option of the holder and mandatorily convertible upon a) a firm-commitment underwritten qualified initial public offering resulting in at least $75.0 million of gross proceeds to the Company (Qualified IPO) or b) the vote or written consent of holders of at least 56% of the outstanding shares of convertible preferred stock (Requisite Holders) into common stock. The conversion ratio of each series of the convertible preferred stock is determined by dividing the original issuance price of each series by the applicable conversion price of each series. The Series A convertible preferred stock conversion price initially is equal to $9.1319, and the Series Seed convertible preferred stock conversion price initially is equal to $0.9105, each subject to appropriate adjustment in the event of any stock dividend, stock split, combination or other similar recapitalization and other adjustments, including adjustment if common stock is issued for less than the original issue price of each series of convertible preferred stock (subject to certain exceptions, including shares of common stock in a Qualified IPO).
Voting
The holder of each share of Series Seed and Series A convertible preferred stock is entitled to one vote for each share of common stock into which such shares of Series Seed and Series A convertible preferred stock could then be converted and votes together with the holders of common stock as a single class, on an as-converted to common stock basis.
Redemption
The holders of Series Seed and Series A convertible preferred stock do not have any redemption rights, except for the contingent redemption upon the occurrence of a Deemed Liquidation Event (as defined below).
Liquidation
In the event of any voluntary or involuntary liquidation, dissolution or winding up of the Company, or upon the occurrence of a Deemed Liquidation Event (as defined below), the holders of the Series Seed and Series A convertible preferred stock are entitled to receive an amount per share equal to the greater of (i) the respective original issuance price, plus any dividends declared but unpaid or (ii) the amount payable with respect to such share if it was converted to common stock immediately prior to such liquidation, dissolution, winding up or Deemed Liquidation Event (as defined below). Liquidation payments to the holders of the Series Seed and Series A convertible preferred stock have priority and
CITY THERAPEUTICS, INC.
NOTES TO CONSOLIDATED FINANCIAL STATEMENTS (Continued)
10. Convertible Preferred Stock (Continued)
are made in preference to any payments to the holders of common stock. After full payment of the liquidation preference to the holders of the Series Seed and Series A convertible preferred stock, the remaining assets, if any, will be distributed to the holders of shares of common stock, pro rata based on the number of shares held by each holder. Unless the Requisite Holders, elect otherwise, a “Deemed Liquidation Event” shall include a merger, consolidation, or share exchange (other than one in which stockholders of the Company own a majority by voting power of the outstanding shares of the surviving or acquiring corporation) or a sale, lease, transfer, exclusive license or other disposition of all or substantially all of the assets of the Company taken as a whole or the sale or disposition of one or more subsidiaries if substantially all of the assets of the Company and its subsidiaries are held by such subsidiary or subsidiaries.
Dividends
The Company shall not declare, pay or set aside any dividends on shares of any class of capital stock of the Company unless the holders of the Series A and Series Seed convertible preferred stock shall first receive, or simultaneously receive, a noncumulative dividend on each outstanding share of the Series A and Series Seed convertible preferred stock equal to 8% of the original issue prices. No such dividends have been declared or paid through December 31, 2025.
11. Common Stock
The Company was authorized to issue 30,000,000 shares of $0.0001 par value common stock as of December 31, 2024 and 2025, respectively.
On April 17, 2024, the Company issued 32,424 shares of common stock to OSIF in conjunction with the OSIF License Agreement (see Note 9). The OSIF common stock was fully vested at the time of issuance.
Features of the Common Stock
The common stock has a par value of $0.0001, and the holders of common stock are entitled to one vote for each share of common stock held at all meetings of stockholders and written actions in lieu of meetings provided. All dividends shall be declared and paid pro rata according to the number of shares held by each stockholder. In the event of a liquidation, dissolution, or winding up of the Company, the common stock ranks behind the preferred stock.
Common stock reserved for future issuance consists of the following:
| |
|
|
As of December 31,
|
|
| |
|
|
2024
|
|
|
2025
|
|
|
Series Seed convertible preferred stock
|
|
|
|
|
4,991,760 |
|
|
|
|
|
4,991,760 |
|
|
|
Series A convertible preferred stock
|
|
|
|
|
15,372,145 |
|
|
|
|
|
15,372,145 |
|
|
|
Unvested restricted stock
|
|
|
|
|
2,020,411 |
|
|
|
|
|
2,081,638 |
|
|
|
Stock options issued and outstanding
|
|
|
|
|
4,468,283 |
|
|
|
|
|
4,768,315 |
|
|
|
Shares available for issuance under the Plan
|
|
|
|
|
552,228 |
|
|
|
|
|
624,683 |
|
|
|
Total
|
|
|
|
|
27,404,827 |
|
|
|
|
|
27,838,541 |
|
|
CITY THERAPEUTICS, INC.
NOTES TO CONSOLIDATED FINANCIAL STATEMENTS (Continued)
12. Stock-based Compensation
2023 Stock Plan
The 2023 Plan was originally adopted with 1,136,740 shares of common stock reserved for issuance. The Board subsequently increased the number of shares reserved under the 2023 Plan at various times during 2024 and 2025. As of December 31, 2024 and 2025, the Company had reserved 5,020,511 and 5,392,981 shares of common stock for issuance under the 2023 Plan, respectively.
The 2023 Plan authorizes the Board or a committee of the Board to grant incentive stock options, nonqualified stock options, and restricted stock awards to eligible employees, directors, and non-employee consultants of the Company. The terms of stock awards agreements, including vesting requirements, are determined by the Board and are subject to the provisions of the 2023 Plan. The stock options granted to employees follow various vesting periods ranging from immediate to four-year vesting and have a term of up to ten years from the date of grant.
As of December 31, 2025, the Company has 624,683 shares of common stock reserved for future issuance under the 2023 Plan. There are no additional shares of common stock available for future issuance outside the common stock reserved under the 2023 Plan.
Stock Option Valuation
The Company estimates the fair value of stock options using the Black-Scholes option-pricing model. This model incorporates various assumptions, including the expected volatility, expected term, and interest rates.
The following table presents, on a weighted-average basis, the assumptions used in the Black-Scholes option-pricing model to determine the fair value of stock options granted during 2024 and 2025:
| |
|
|
Years ended December 31,
|
|
| |
|
|
2024
|
|
|
2025
|
|
|
Risk-free interest rate
|
|
|
3.55% – 4.47%
|
|
|
3.67% – 4.22%
|
|
|
Dividend yield
|
|
|
0%
|
|
|
0%
|
|
|
Expected term of options (years) range
|
|
|
5.00 – 6.08
|
|
|
5.27 – 6.08
|
|
|
Volatility rate range
|
|
|
67.68% – 69.21%
|
|
|
87.73% – 111.06%
|
|
The weighted-average grant-date fair value of stock options granted during the years ended December 31, 2024 and 2025 was $3.11 and $1.87 per share, respectively.
CITY THERAPEUTICS, INC.
NOTES TO CONSOLIDATED FINANCIAL STATEMENTS (Continued)
12. Stock-based Compensation (Continued)
Time-Based Stock Options
The following table summarizes the Company’s time-based stock option activity for the year ended December 31, 2025:
| |
|
|
Number of
Outstanding
Options
|
|
|
Weighted
Average
Exercise
Price
|
|
|
Weighted
Average
Remaining
Contractual
Term
(in years)
|
|
|
Aggregate
Intrinsic
Value
|
|
|
Balance at December 31, 2024
|
|
|
|
|
3,153,342 |
|
|
|
|
$ |
0.83 |
|
|
|
|
|
9.67 |
|
|
|
|
$ |
9,508 |
|
|
|
Granted
|
|
|
|
|
1,130,466 |
|
|
|
|
|
1.65 |
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
Exercised
|
|
|
|
|
(896,864) |
|
|
|
|
|
0.71 |
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
Cancelled
|
|
|
|
|
(10,983) |
|
|
|
|
|
1.84 |
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
Forfeited
|
|
|
|
|
(8,093) |
|
|
|
|
|
0.92 |
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
Balance at December 31, 2025
|
|
|
|
|
3,367,868 |
|
|
|
|
$ |
1.13 |
|
|
|
|
|
8.98 |
|
|
|
|
$ |
2,383 |
|
|
|
Vested and expected to vest as of December 31, 2025
|
|
|
|
|
3,367,868 |
|
|
|
|
$ |
1.13 |
|
|
|
|
|
8.98 |
|
|
|
|
$ |
2,383 |
|
|
|
Vested and exercisable at December 31, 2025
|
|
|
|
|
847,617 |
|
|
|
|
$ |
0.65 |
|
|
|
|
|
8.58 |
|
|
|
|
$ |
1,350 |
|
|
As of December 31, 2025, there was $6.4 million of total unrecognized compensation cost related to time-based unvested stock options, and the Company expects to recognize such amount over a remaining weighted-average period of 2.9 years.
Performance-Based Stock Options
The Company has granted stock options to certain employees to purchase shares of common stock that contain performance-based vesting criteria related to corporate or scientific milestones. The fair value of each option grant with performance-based vesting was estimated on the date of grant. Recognition of stock-based compensation expense associated with these performance-based stock options commences when the performance condition is considered probable of achievement, using management’s best estimates, which consider the inherent risk and uncertainty regarding the future outcomes of the milestones. As of December 31, 2025, no stock-based compensation expense has been recognized for performance-based stock options as the achievement of the underlying performance conditions was not considered probable.
CITY THERAPEUTICS, INC.
NOTES TO CONSOLIDATED FINANCIAL STATEMENTS (Continued)
12. Stock-based Compensation (Continued)
The following table summarizes the Company’s performance-based stock option activity for the year ended December 31, 2025:
| |
|
|
Number of
Outstanding
Options
|
|
|
Weighted
Average
Exercise
Price
|
|
|
Weighted
Average
Remaining
Contractual
Term
(in years)
|
|
|
Aggregate
Intrinsic
Value
|
|
|
Balance at December 31, 2024
|
|
|
|
|
1,314,941 |
|
|
|
|
$ |
0.85 |
|
|
|
|
|
9.73 |
|
|
|
|
$ |
3,929 |
|
|
|
Granted
|
|
|
|
|
463,605 |
|
|
|
|
|
1.56 |
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
Exercised
|
|
|
|
|
(378,099) |
|
|
|
|
|
0.71 |
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
Balance at December 31, 2025
|
|
|
|
|
1,400,447 |
|
|
|
|
$ |
1.13 |
|
|
|
|
|
9.02 |
|
|
|
|
$ |
1,000 |
|
|
|
Vested and exercisable at December 31, 2025
|
|
|
|
|
— |
|
|
|
|
$ |
— |
|
|
|
|
|
— |
|
|
|
|
$ |
— |
|
|
As of December 31, 2025, there was $4.1 million of total unrecognized compensation cost related to performance-based unvested stock options, and the Company expects to recognize such amount when the achievement of the underlying performance conditions become probable.
Restricted Common Stock Awards
The Company awarded restricted common stock to employees and non-employees under its 2023 Plan and may continue to award restricted common stock to employees and non-employees under the 2023 Plan. Additionally, employees who exercise their option awards prior to the vesting of that award hold common stock that is subject to further time or performance based vesting conditions and therefore classified as restricted stock for accounting purposes. The fair value of each share of restricted common stock is based on the market price of the Company’s common stock on the date of grant.
Time-Based Restricted Common Stock Awards
Restricted common stock that have time-based vesting conditions vest over a four-year period, subject to the employee’s continued employment with, or service to, the Company on each vesting date. Compensation expense is recognized on a straight-line basis over the vesting period.
The following table summarizes the Company’s time-based restricted common stock award activity for the year ended December 31, 2025:
| |
|
|
Number
of Shares
|
|
|
Weighted
Average
Grant Date
Fair Value
|
|
|
Unvested restricted common stock at December 31, 2024
|
|
|
|
|
2,020,411 |
|
|
|
|
$ |
0.10 |
|
|
|
Issued
|
|
|
|
|
772,539 |
|
|
|
|
|
3.27 |
|
|
|
Vested
|
|
|
|
|
(1,089,411) |
|
|
|
|
|
1.10 |
|
|
|
Unvested restricted common stock at December 31, 2025
|
|
|
|
|
1,703,539 |
|
|
|
|
$ |
1.02 |
|
|
As of December 31, 2025, there was $1.3 million of total unrecognized compensation cost related to time-based restricted common stock, and the Company expects to recognize such amount over a remaining weighted-average period of 2.3 years.
CITY THERAPEUTICS, INC.
NOTES TO CONSOLIDATED FINANCIAL STATEMENTS (Continued)
12. Stock-based Compensation (Continued)
Performance-Based Restricted Common Stock Awards
Restricted common stock that have performance-based vesting conditions vest when certain milestones are achieved and approved by the Board, subject to the employee’s continued employment with, or service to, the Company on each vesting date. Compensation expense is recognized upon the achievement of the certain milestones.
The following table summarizes the Company’s performance-based restricted common stock award activity for the year ended December 31, 2025:
| |
|
|
Number
of Shares
|
|
|
Weighted
Average
Grant Date
Fair Value
|
|
|
Unvested restricted common stock at December 31, 2024
|
|
|
|
|
— |
|
|
|
|
$ |
— |
|
|
|
Issued
|
|
|
|
|
378,099 |
|
|
|
|
|
3.65 |
|
|
|
Unvested restricted common stock at December 31, 2025
|
|
|
|
|
378,099 |
|
|
|
|
$ |
3.65 |
|
|
As of December 31, 2025, there was $1.1 million of total unrecognized compensation cost related to performance-based restricted common stock, and the Company expects to recognize such amount when the achievement of the underlying performance conditions become probable.
Stock-Based Compensation
The following table below summarizes the classification of the Company’s stock-based compensation expense related to stock options and restricted common stock awards in the consolidated statements of operations and comprehensive loss (in thousands):
| |
|
|
Years ended December 31,
|
|
| |
|
|
2024
|
|
|
2025
|
|
|
Research and development
|
|
|
|
$ |
445 |
|
|
|
|
$ |
955 |
|
|
|
General and administrative
|
|
|
|
|
560 |
|
|
|
|
|
2,042 |
|
|
|
Total stock-based compensation expense
|
|
|
|
$ |
1,005 |
|
|
|
|
$ |
2,997 |
|
|
13. 401(k) Plan
The Company has a defined-contribution savings plan under Section 401(k) of the IRC (401(k) Plan). The 401(k) Plan covers all employees who meet defined minimum age and service requirements and allows participants to defer a portion of their annual compensation on a pretax basis. The Company will make a contribution of 4% of each employee’s annual compensation up to annual safe harbor limits. The Company has expensed $0.1 million and $0.4 million in 401(k) contributions for the years ended December 31, 2024 and 2025, respectively.
14. Income Taxes
For the years ended December 31, 2024 and 2025, the Company did not recognize any current or deferred income tax provision or benefit and is in a full valuation allowance. The Company did not make any federal, state, local or foreign income tax payments, net of refunds received, during the years ended December 31, 2024 and 2025.
CITY THERAPEUTICS, INC.
NOTES TO CONSOLIDATED FINANCIAL STATEMENTS (Continued)
14. Income Taxes (Continued)
The components of the Company’s loss before income taxes are as follows (in thousands):
| |
|
|
Years ended December 31,
|
|
| |
|
|
2024
|
|
|
2025
|
|
|
U.S. Domestic
|
|
|
|
$ |
(27,836) |
|
|
|
|
$ |
(58,505) |
|
|
|
Foreign
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
Pretax loss from operations
|
|
|
|
$ |
(27,836) |
|
|
|
|
$ |
(58,505) |
|
|
A reconciliation of income tax expense computed at the statutory corporate income tax rate to the effective income tax rate after the adoption of ASU 2023-09 for the years ended December 31, 2024 and 2025 is as follows (in thousands):
| |
|
|
Year Ended December 31,
|
|
|
Year Ended December 31,
|
|
| |
|
|
2024
|
|
|
2025
|
|
| |
|
|
Amount
|
|
|
Percent
|
|
|
Amount
|
|
|
Percent
|
|
|
U.S. federal statutory tax rate
|
|
|
|
$ |
(5,845) |
|
|
|
|
|
21.0% |
|
|
|
|
$ |
(12,286) |
|
|
|
|
|
21.0% |
|
|
|
Tax credits
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
Research and development tax credits
|
|
|
|
|
(798) |
|
|
|
|
|
2.9 |
|
|
|
|
|
(1,221) |
|
|
|
|
|
2.1 |
|
|
|
Changes in valuation allowances
|
|
|
|
|
6,154 |
|
|
|
|
|
(22.1) |
|
|
|
|
|
12,295 |
|
|
|
|
|
(21.0) |
|
|
|
Nontaxable or nondeductible items
|
|
|
|
|
489 |
|
|
|
|
|
(1.8) |
|
|
|
|
|
1,222 |
|
|
|
|
|
(2.1) |
|
|
|
Other reconciling items
|
|
|
|
|
— |
|
|
|
|
|
0.0 |
|
|
|
|
|
(10) |
|
|
|
|
|
0.0 |
|
|
|
Income tax expense and effective tax
rate
|
|
|
|
$ |
— |
|
|
|
|
|
0.0% |
|
|
|
|
$ |
— |
|
|
|
|
|
0.0% |
|
|
The effective income tax rate differs from the U.S. federal statutory rate of 21.0% primarily due to the valuation allowance maintained against the Company’s net deferred tax assets.
The significant components of the Company’s deferred tax assets and liabilities as of December 31, 2024 and 2025 are comprised of the following (in thousands):
| |
|
|
Years ended December 31,
|
|
| |
|
|
2024
|
|
|
2025
|
|
|
Deferred tax assets:
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
Net operating loss carryforwards
|
|
|
|
$ |
797 |
|
|
|
|
$ |
13,001 |
|
|
|
Tax credit carryforwards
|
|
|
|
|
1,118 |
|
|
|
|
|
2,987 |
|
|
|
Operating lease liability
|
|
|
|
|
3,253 |
|
|
|
|
|
6,944 |
|
|
|
Deferred revenue
|
|
|
|
|
— |
|
|
|
|
|
463 |
|
|
|
Amortization – start up costs
|
|
|
|
|
65 |
|
|
|
|
|
61 |
|
|
|
Capitalized research and development costs
|
|
|
|
|
6,437 |
|
|
|
|
|
7,964 |
|
|
|
Accrued expenses and other
|
|
|
|
|
274 |
|
|
|
|
|
1,101 |
|
|
|
Total deferred tax assets
|
|
|
|
|
11,944 |
|
|
|
|
|
32,521 |
|
|
|
Valuation allowance
|
|
|
|
|
(8,733) |
|
|
|
|
|
(25,240) |
|
|
|
Net deferred tax assets
|
|
|
|
|
3,211 |
|
|
|
|
|
7,281 |
|
|
|
Deferred tax liabilities:
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
Operating lease right-of-use asset
|
|
|
|
|
(2,846) |
|
|
|
|
|
(6,791) |
|
|
|
Depreciation
|
|
|
|
|
(365) |
|
|
|
|
|
(490) |
|
|
|
Total deferred tax liabilities
|
|
|
|
|
(3,211) |
|
|
|
|
|
(7,281) |
|
|
|
Net deferred tax assets
|
|
|
|
$ |
— |
|
|
|
|
$ |
— |
|
|
CITY THERAPEUTICS, INC.
NOTES TO CONSOLIDATED FINANCIAL STATEMENTS (Continued)
14. Income Taxes (Continued)
The Company has evaluated the positive and negative evidence bearing upon its ability to realize its deferred tax assets. Management has considered the Company’s history of cumulative net losses, estimated future taxable income, and prudent and feasible tax planning strategies and has concluded that it is more likely than not that the Company will not realize the benefits of its net deferred tax assets. Accordingly, a full valuation allowance has been established against these net deferred tax assets as of December 31, 2024 and 2025. The Company’s valuation allowance increased by approximately $16.5 million during the year ended December 31, 2025, primarily due to the increase in net operating loss carryforwards and capitalization of research and experimental expenditures.
The Company’s change in its valuation allowance account with respect to deferred tax assets is as follows (in thousands):
| |
|
|
Year ended December 31,
|
|
| |
|
|
2024
|
|
|
2025
|
|
|
Beginning Balance
|
|
|
|
$ |
721 |
|
|
|
|
$ |
8,733 |
|
|
|
Additions
|
|
|
|
|
8,012 |
|
|
|
|
|
16,507 |
|
|
|
Reductions
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
Ending Balance
|
|
|
|
$ |
8,733 |
|
|
|
|
$ |
25,240 |
|
|
As of December 31, 2024 and 2025, the Company had U.S. federal net operating loss (NOL) carryforwards of approximately $3.1 million and $47.1 million, respectively, which do not expire and may be available to offset future U.S. federal taxable income. U.S. federal NOL carryforwards generated after 2017 can be carried over indefinitely but will generally limit the NOL deduction to the lesser of the NOL carryover or 80% of a corporation’s taxable income, as required under Section 172 of the Internal Revenue Code (IRC).
As of December 31, 2024 and 2025, the Company also had U.S. state NOL carryforwards of approximately $2.5 million and $49.2 million, respectively, which expire at various dates through 2045 and may be available to offset future U.S. state taxable income.
As of December 31, 2024 and 2025, the Company had U.S. federal research and development (R&D) tax credit carryforwards of approximately $0.8 million and $2.0 million, respectively, which expire at various dates through 2045 and may be available to offset future U.S. federal income tax liabilities.
As of December 31, 2024 and 2025, the Company had U.S. state R&D tax credit carryforwards of approximately $0.4 million and $1.2 million, respectively, which expire at various dates through 2040 and may be available to reduce future U.S. state income tax liabilities.
Under the provisions of the IRC, the NOL and tax credit carryforwards are subject to review and possible adjustment by the Internal Revenue Service and state tax authorities. Utilization of U.S. federal and state NOL and tax credit carryforwards may be subject to a substantial annual limitation under Section 382 and Section 383 of the IRC, and corresponding provisions of state law, due to ownership changes that have occurred previously or that could occur in the future. These ownership changes may limit the amount of U.S. federal and state NOL and tax credit carryforwards that can be utilized annually to offset future taxable income and tax liabilities, respectively. The Company has not yet conducted a comprehensive study to assess whether any changes of ownership have occurred since its formation. Any limitation may result in expiration of a portion of the U.S. federal and state NOL carryforward or tax credit carryforwards before utilization, which would be offset by a change in the Company’s valuation allowance.
The Company has not yet conducted an R&D tax credit study. Such a study, once undertaken by the Company, may result in an adjustment to the R&D tax credit carryforwards. A full valuation allowance has
CITY THERAPEUTICS, INC.
NOTES TO CONSOLIDATED FINANCIAL STATEMENTS (Continued)
14. Income Taxes (Continued)
been provided against the Company’s R&D tax credits and, if an adjustment is required, this adjustment would be offset by an adjustment to the valuation allowance. Thus, there would be no impact to the consolidated balance sheets or consolidated statements of operations and comprehensive loss if an adjustment is required.
The Company does not have any unrecognized tax benefit and therefore, no amounts are reflected in the determination of the Company’s deferred tax assets. If there were unrecognized tax benefits to be recognized, none of these amounts would affect the Company’s effective tax rate, since it would be offset by an equal corresponding adjustment in the deferred tax asset valuation allowance. The Company assesses the uncertainty in its income tax positions to determine whether a tax position of the Company is more likely than not to be sustained upon examination, including resolution of any related appeals of litigation processes, based on the technical merits of the position. For tax positions meeting the more-likely-than-not threshold, the tax amount recognized in the financial statements is reduced by the largest benefit that has a greater than 50% likelihood of being realized upon the ultimate settlement with the relevant taxing authority. The Company will recognize interest and/or penalties related to uncertain tax benefits in income tax expense as they arise. As of December 31, 2024 and 2025, the Company had no reserves for uncertain tax benefits or related interest or penalties.
The Company files tax returns as prescribed by the tax laws of the jurisdictions in which it operates. In a normal course of business, the Company is subject to examination by U.S. federal and state as well as foreign jurisdictions, where applicable. The Company’s tax years are still open since inception. To the extent that the Company has tax attribute carryforwards, the tax year in which the attributes were generated may still be adjusted upon examination by the U.S. Internal Revenue Service or state tax authorities to the extent utilized in a future period. The Company is not currently under examination by any tax authorities.
On July 4, 2025, the U.S. government enacted The One Big Beautiful Bill Act of 2025 (OBBBA) which includes, among other provisions, changes to the U.S. corporate income tax system. In accordance with ASC 740, the Company recognized the effects of the new tax law in the period that includes the enactment date. While OBBBA did not have a material impact on the Company’s consolidated financial statements as of and for the years ended December 31, 2024 and 2025, the Company will continue to assess the changes. However, the Company does not expect such changes will have a material effect on its consolidated financial statements in future periods.
15. Segments
The Company’s operations are all in the United States. The Company operates and manages its business as a single operating and reportable segment. Factors considered in determining operating and reportable segments include the organization of the business, the nature of technology and the information reviewed by the Company’s CODM. The Company’s CODM is our CEO.
Operating segments are defined as components of an enterprise for which discrete financial information is available and is evaluated regularly by the CODM on a consolidated basis, in deciding how to allocate resources and assess performance. The Company’s revenues are derived from research collaboration and license agreements, which are further described in Note 9 — Significant Agreements. Consolidated net loss is used by the CODM to make key operating decisions, such as the determination of the rate at which the Company seeks to grow its research and development initiatives and the allocation of capital between research and development and general and administrative expenses. The CODM does not receive asset information other than what is presented on the consolidated balance sheets.
CITY THERAPEUTICS, INC.
NOTES TO CONSOLIDATED FINANCIAL STATEMENTS (Continued)
15. Segments (Continued)
The following table presents financial information, including significant segment expenses, which are regularly provided to the CODM and included within consolidated net loss (in thousands):
| |
|
|
Years Ended December 31,
|
|
| |
|
|
2024
|
|
|
2025
|
|
|
Collaboration revenue:
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
Ocular (Bausch + Lomb)
|
|
|
|
$ |
2,778 |
|
|
|
|
$ |
7,123 |
|
|
|
CNS (Biogen)
|
|
|
|
|
— |
|
|
|
|
|
6,439 |
|
|
|
Total collaboration revenue
|
|
|
|
|
2,778 |
|
|
|
|
|
13,562 |
|
|
|
Research and development expenses:
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
CITY-FXI
|
|
|
|
|
5,694 |
|
|
|
|
|
12,161 |
|
|
|
CITY-RBP4
|
|
|
|
|
2,981 |
|
|
|
|
|
7,109 |
|
|
|
CITY-TFR2
|
|
|
|
|
1,104 |
|
|
|
|
|
2,985 |
|
|
|
Ocular
|
|
|
|
|
919 |
|
|
|
|
|
2,563 |
|
|
|
CNS
|
|
|
|
|
240 |
|
|
|
|
|
2,711 |
|
|
|
Personnel-related research and development expenses (including stock-based compensation expense of $445 and $955 for 2024 and 2025, respectively)
|
|
|
|
|
5,383 |
|
|
|
|
|
11,453 |
|
|
|
Other indirect research and development expenses(1)
|
|
|
|
|
9,361 |
|
|
|
|
|
19,438 |
|
|
|
Total research and development expenses
|
|
|
|
|
25,682 |
|
|
|
|
|
58,420 |
|
|
|
General and administrative expenses:
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
Personnel-related expenses (including stock-based compensation expense of $560 and $2,042 for 2024 and 2025, respectively)
|
|
|
|
|
1,757 |
|
|
|
|
|
5,949 |
|
|
|
Other general and administrative expenses(2)
|
|
|
|
|
4,429 |
|
|
|
|
|
9,115 |
|
|
|
Total general and administrative expenses
|
|
|
|
|
6,186 |
|
|
|
|
|
15,064 |
|
|
|
Other segment items, net(3)
|
|
|
|
|
1,254 |
|
|
|
|
|
1,417 |
|
|
|
Net loss and comprehensive loss
|
|
|
|
$ |
(27,836) |
|
|
|
|
$ |
(58,505) |
|
|
(1)
Other indirect research and development expenses consist of platform research costs, and laboratory supplies, equipment, facility-related expenses and other general research costs that are not specifically attributable to an individual research program.
(2)
Other general and administrative expenses consist of professional fees, facilities, depreciation, information technology and other general and administrative costs that are not personnel-related.
(3)
Other segment items, net consists of interest income, interest expense and other expense, net.
16. Net Loss Per Share
Under the two-class method, basic net loss per share is calculated by dividing the net loss attributable to common stockholders by the weighted average number of shares of common stock outstanding during the period, without consideration for potentially dilutive securities. The net loss available to common stockholders was not allocated to the convertible preferred stock as the holders did not have a contractual obligation to share in losses. Diluted net loss per share is the same as basic net loss per share for the years presented since the effects of any potentially dilutive securities, would be antidilutive given the net loss of the Company.
CITY THERAPEUTICS, INC.
NOTES TO CONSOLIDATED FINANCIAL STATEMENTS (Continued)
16. Net Loss Per Share (Continued)
| |
|
|
Years Ended December 31,
|
|
| |
|
|
2024
|
|
|
2025
|
|
|
Numerator:
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
Net loss attributable to common stockholders – basic and diluted
|
|
|
|
$ |
(27,836) |
|
|
|
|
$ |
(58,505) |
|
|
|
Denominator:
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
Weighted-average common shares outstanding – basic and diluted
|
|
|
|
|
3,738,335 |
|
|
|
|
|
5,157,781 |
|
|
|
Net loss per common share – basic and diluted
|
|
|
|
$ |
(7.45) |
|
|
|
|
$ |
(11.34) |
|
|
For accounting purposes, the computation of basic and diluted weighted-average common shares outstanding for the years ended December 31, 2024 and 2025, excludes all shares of unvested restricted common stock as such shares are not considered outstanding. See Note 12 — Stock-Based Compensation, for more information.
The following outstanding potentially dilutive securities have been excluded from the calculation of diluted net loss per share attributable to common stockholders, as their effect is antidilutive:
| |
|
|
Years ended December 31,
|
|
| |
|
|
2024
|
|
|
2025
|
|
|
Series Seed convertible preferred stock (as-converted basis)
|
|
|
|
|
4,991,760 |
|
|
|
|
|
4,991,760 |
|
|
|
Series A convertible preferred stock (as-converted basis)
|
|
|
|
|
15,372,145 |
|
|
|
|
|
15,372,145 |
|
|
|
Unvested restricted stock
|
|
|
|
|
2,020,411 |
|
|
|
|
|
2,081,638 |
|
|
|
Stock options issued and outstanding
|
|
|
|
|
4,468,283 |
|
|
|
|
|
4,768,315 |
|
|
|
Total
|
|
|
|
|
26,852,599 |
|
|
|
|
|
27,213,858 |
|
|
The potentially dilutive securities related to the Biogen Note and B+L Note were excluded from the table above for the years ended December 31, 2024 and 2025, as applicable, as the number of shares that would be issuable upon conversion was indeterminable. See Note 5 — Convertible Notes, for more information.
17. Related Party Transactions
Alexandria Real Estate Equities, Inc.
For the years ended December 31, 2024 and 2025, the Company paid $4.2 million and $6.2 million, respectively, in rent and facility-related fees to a subsidiary of Alexandria Real Estate Equities, Inc., a related party, in connection with leasing its headquarters at 399 Binney Street (see Note 7). No payments were made to Alexandria Venture Investments, LLC, an investor in the Company, holding shares of Series A convertible preferred stock as of December 31, 2024 and 2025.
Company Founder Consulting Agreement & Convertible Note
During the years ended December 31, 2024 and 2025, the Company made payments of $0.2 million and $0.2 million, respectively, to a Founder, who is a member of the Company’s Board, for consulting services.
In September 2023, in connection with the Series Seed convertible preferred stock closing, the Company issued and sold 500,000 shares of Series Seed convertible preferred stock to the Founder for an aggregate purchase price of $0.5 million.
CITY THERAPEUTICS, INC.
NOTES TO CONSOLIDATED FINANCIAL STATEMENTS (Continued)
17. Related Party Transactions (Continued)
In January and February 2024, the Company issued convertible notes with an aggregate principal amount of $0.5 million to the Founder, who is a member of the Company’s Board (see Note 5). The convertible notes converted into 56,095 shares of Series A convertible preferred stock in connection with the closing, at a discounted conversion price of $9.02655 per share. This price reflects a 10% discount to the $10.0295 per share purchase price paid by new investors in the Series A financing.
ARCH Venture Fund XII L.P.
In January and February 2024, the Company issued convertible notes with an aggregate principal amount of $4.5 million to ARCH, an investor in the Company’s convertible preferred stock financings (see Note 5). The convertible notes converted into 504,869 shares of Series A convertible preferred stock in connection with the closing, at a discounted conversion price of $9.02655 per share. This price reflects a 10% discount to the $10.0295 per share purchase price paid by new investors in the Series A convertible preferred stock financing.
18. Subsequent Events
The Company has evaluated subsequent events from the balance sheet date through July 14, 2026, the date at which the consolidated financial statements were available to be issued and, only with respect to the stock split, through October 9, 2026, the date the consolidated financial statements were available to be reissued. The Company has determined there are no events subject to additional disclosure, other than those disclosed below.
Option Grants
During the period from January 2026 through July 2026, the Company granted an aggregate of 2,904,402 stock options to employees, directors, and non-employee consultants under the 2023 Plan with exercise prices ranging from $1.84 to $7.89. The Company also granted an aggregate of 109,828 shares of restricted common stock to non-employee consultants under the 2023 Plan. The Company is evaluating the accounting impact of these grants.
Performance-Based Stock Options Milestone Achievement
In January 2026, the Company determined that the achievement of a scientific milestone was probable, triggering the vesting condition on certain performance-based stock options. As a result, the Company recognized $3.4 million of stock-based compensation expense related to the vesting of 1,071,044 stock options.
Amendment to 2023 Plan
During January 2026, May 2026 and June 2026, the Board amended the 2023 Plan to increase the number of shares reserved under the 2023 Plan to 7,985,921, 8,535,071, and 10,017,776, respectively.
Series B Convertible Preferred Stock Financing
Beginning in May 2026 and in multiple closings through June 2026, the Company issued shares of its Series B convertible preferred stock pursuant to a Series B Convertible Preferred Stock Purchase Agreement (Series B Agreement) with certain investors. Pursuant to the Series B Agreement, the Company issued an aggregate of 7,725,348 shares of Series B convertible preferred stock. Of the 7,725,348 shares issued, 6,801,318 shares were issued at a price per share of $14.6295 for aggregate gross proceeds of $99.5 million. Additionally, the Company’s outstanding B+L Note and related accrued interest, at the time of the Series B Agreement, totaling $11.5 million, converted into 924,030 shares of Series B convertible preferred stock at a price per share of $12.4351.
CITY THERAPEUTICS, INC.
NOTES TO CONSOLIDATED FINANCIAL STATEMENTS (Continued)
18. Subsequent Events (Continued)
The Company is continuing to evaluate the accounting impact of the Series B convertible preferred financing, including the conversion of the outstanding convertible note, and will reflect the completed accounting impact in its financial statements for the period ended June 30, 2026.
Stock Split
On October 8, 2026, the Company effected a 1.0983-for-one stock split of its issued and outstanding shares of common stock and a proportional adjustment to the existing conversion ratios of each series of the Company’s convertible preferred stock (see Note 10). Accordingly, all share and per share amounts for all periods presented in the accompanying consolidated financial statements and notes thereto have been adjusted retroactively, where applicable, to reflect this stock split and adjustment of the preferred stock conversion ratios.
Events subsequent to the original issuance of consolidated financial statements (unaudited)
In connection with the reissuance of the financial statements, the Company has evaluated subsequent events through October 9, 2026, the date the financial statements were available to be reissued.
2026 Stock Option and Incentive Plan
On September 19, 2026, the Company’s Board adopted, and on October 8, 2026, the Company’s stockholders approved, the 2026 Stock Option and Incentive Plan (2026 Plan), which will become effective on the date immediately prior to the effectiveness of the registration statement for the Company's IPO. The 2026 Plan provides for the grant of incentive stock options, nonqualified stock options, restricted stock awards, and other stock-based awards. A total of 6,516,101 shares of common stock were initially reserved for issuance under the 2026 Plan. The number of shares reserved and available for issuance under the 2026 Plan will automatically increase on January 1, 2027 and each January thereafter during the term of the 2026 Plan, by 5% of the outstanding number of shares of common stock on the immediately preceding December 31 (including, for this purpose, shares underlying any outstanding prefunded warrants) or such lesser number of shares as determined by the administrator of the 2026 Plan. Upon the effectiveness of the 2026 Plan, no further grants will be made under the 2023 Plan. The shares of common stock underlying any awards under the 2026 Plan or the 2023 Plan that are forfeited, cancelled, held back upon exercise or settlement of an award to satisfy the exercise price or tax withholding, reacquired by the Company prior to vesting, satisfied without the issuance of stock, or are otherwise terminated (other than by exercise) will be added back to the shares of common stock available for issuance under the 2026 Plan. Upon the effectiveness of the 2026 Plan, no further grants will be made under the 2023 Plan.
2026 Employee Stock Purchase Plan
On September 19, 2026, the Company's Board adopted, and on October 8, 2026, the Company's stockholders approved, the 2026 Employee Stock Purchase Plan (ESPP), which will become effective on the date immediately prior to the effectiveness of the registration statement for the Company’s IPO. A total of 503,322 shares of common stock were initially reserved for issuance under the ESPP. The ESPP provides that the number of shares reserved and available for issuance will automatically increase on January 1, 2027 and each January 1 thereafter through January 1, 2036, by the least of (i) 1% of the outstanding number of shares of common stock on the immediately preceding December 31 (including, for this purpose, shares underlying any outstanding prefunded warrants), (ii) 1,509,966 shares of common stock, or (iii) such number of shares of common stock as determined by the administrator of the ESPP.
CITY THERAPEUTICS, INC.
CONDENSED CONSOLIDATED BALANCE SHEETS
(UNAUDITED, IN THOUSANDS, EXCEPT SHARE AND PER SHARE AMOUNTS)
| |
|
|
December 31,
2025
|
|
|
June 30,
2026
|
|
|
Assets
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
Current assets
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
Cash and cash equivalents
|
|
|
|
$ |
112,793 |
|
|
|
|
$ |
169,605 |
|
|
|
Accounts receivable
|
|
|
|
|
1,209 |
|
|
|
|
|
5,119 |
|
|
|
Prepaid expenses and other current assets
|
|
|
|
|
2,871 |
|
|
|
|
|
964 |
|
|
|
Total current assets
|
|
|
|
|
116,873 |
|
|
|
|
|
175,688 |
|
|
|
Property and equipment, net
|
|
|
|
|
5,489 |
|
|
|
|
|
6,169 |
|
|
|
Operating lease right-of-use asset
|
|
|
|
|
24,857 |
|
|
|
|
|
23,321 |
|
|
|
Restricted cash
|
|
|
|
|
2,504 |
|
|
|
|
|
2,504 |
|
|
|
Deferred offering costs
|
|
|
|
|
— |
|
|
|
|
|
895 |
|
|
|
Other non-current assets
|
|
|
|
|
45 |
|
|
|
|
|
34 |
|
|
|
Total assets
|
|
|
|
$ |
149,768 |
|
|
|
|
$ |
208,611 |
|
|
|
Liabilities, convertible preferred stock and stockholders’ deficit
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
Current liabilities
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
Accounts payable
|
|
|
|
$ |
2,695 |
|
|
|
|
$ |
5,105 |
|
|
|
Convertible notes, current
|
|
|
|
|
10,805 |
|
|
|
|
|
33,089 |
|
|
|
Operating lease liability, current
|
|
|
|
|
5,075 |
|
|
|
|
|
5,126 |
|
|
|
Deferred revenue, current
|
|
|
|
|
11,255 |
|
|
|
|
|
10,791 |
|
|
|
Accrued interest
|
|
|
|
|
2,642 |
|
|
|
|
|
2,647 |
|
|
|
Accrued expenses and other current liabilities
|
|
|
|
|
5,570 |
|
|
|
|
|
7,999 |
|
|
|
Total current liabilities
|
|
|
|
|
38,042 |
|
|
|
|
|
64,757 |
|
|
|
Operating lease liability, non-current
|
|
|
|
|
20,341 |
|
|
|
|
|
18,822 |
|
|
|
Convertible notes, non-current, net
|
|
|
|
|
31,063 |
|
|
|
|
|
— |
|
|
|
Other liabilities, non-current
|
|
|
|
|
463 |
|
|
|
|
|
693 |
|
|
|
Total liabilities
|
|
|
|
|
89,909 |
|
|
|
|
|
84,272 |
|
|
|
Commitments and contingencies (Note 8)
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
Series Seed convertible preferred stock, $0.0001 par value; 4,544,990 and 4,544,990 shares authorized as of December 31, 2025 and June 30, 2026; 4,544,990 and 4,544,990 shares issued and outstanding as of December 31, 2025 and June 30, 2026; aggregate liquidation preference of $4,545 and $4,545 as of December 31, 2025 and June 30, 2026
|
|
|
|
|
4,475 |
|
|
|
|
|
4,475 |
|
|
|
Series A convertible preferred stock, $0.0001 par value; 14,295,436 and 13,996,318
shares authorized as of December 31, 2025 and June 30, 2026; 13,996,318 and
13,996,318 shares issued and outstanding as of December 31, 2025 and
June 30, 2026; aggregate liquidation preference of $140,376 and $140,376 as of
December 31, 2025 and June 30, 2026
|
|
|
|
|
140,039 |
|
|
|
|
|
140,039 |
|
|
|
Series B and B-1 convertible preferred stock, $0.0001 par value; no shares authorized, issued, or outstanding as of December 31, 2025; 7,725,348 shares authorized, issued, and outstanding as of June 30, 2026; aggregate liquidation preference of $0 and $110,990 as of December 31, 2025 and June 30, 2026
|
|
|
|
|
— |
|
|
|
|
|
111,542 |
|
|
|
Stockholders’ deficit:
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
Common stock, $0.0001 par value; 30,000,000 and 42,350,000 shares authorized
as of December 31, 2025 and June 30, 2026; 7,772,189 and 9,627,628 shares
issued as of December 31, 2025 and June 30, 2026; 5,690,551 and 7,663,207
shares outstanding as of December 31, 2025 and June 30, 2026
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
Additional paid-in capital
|
|
|
|
|
4,449 |
|
|
|
|
|
10,830 |
|
|
|
Accumulated deficit
|
|
|
|
|
(89,104) |
|
|
|
|
|
(142,547) |
|
|
|
Total stockholders’ deficit
|
|
|
|
|
(84,655) |
|
|
|
|
|
(131,717) |
|
|
|
Total liabilities, convertible preferred stock and stockholders’ deficit
|
|
|
|
$ |
149,768 |
|
|
|
|
$ |
208,611 |
|
|
The accompanying notes are an integral part of these condensed consolidated financial statements
CITY THERAPEUTICS, INC.
CONDENSED CONSOLIDATED STATEMENTS OF OPERATIONS AND COMPREHENSIVE LOSS
(UNAUDITED, IN THOUSANDS, EXCEPT SHARE AND PER SHARE AMOUNTS)
| |
|
|
Six months ended June 30,
|
|
| |
|
|
2025
|
|
|
2026
|
|
|
Collaboration revenue
|
|
|
|
$ |
5,578 |
|
|
|
|
$ |
6,358 |
|
|
|
Operating expenses:
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
Research and development(1)
|
|
|
|
|
24,375 |
|
|
|
|
|
46,319 |
|
|
|
General and administrative(2)
|
|
|
|
|
7,796 |
|
|
|
|
|
11,170 |
|
|
|
Total operating expenses
|
|
|
|
|
32,171 |
|
|
|
|
|
57,489 |
|
|
|
Loss from operations
|
|
|
|
|
(26,593) |
|
|
|
|
|
(51,131) |
|
|
|
Other income (expense):
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
Interest income
|
|
|
|
|
2,064 |
|
|
|
|
|
1,595 |
|
|
|
Interest expense
|
|
|
|
|
(646) |
|
|
|
|
|
(1,496) |
|
|
|
Other income (expense), net
|
|
|
|
|
893 |
|
|
|
|
|
(2,411) |
|
|
|
Total other income (expense), net
|
|
|
|
|
2,311 |
|
|
|
|
|
(2,312) |
|
|
|
Net loss and comprehensive loss
|
|
|
|
$ |
(24,282) |
|
|
|
|
$ |
(53,443) |
|
|
|
Net loss per share, basic and diluted
|
|
|
|
$ |
(5.02) |
|
|
|
|
$ |
(7.53) |
|
|
|
Weighted-average common shares outstanding, basic and diluted
|
|
|
|
|
4,832,880 |
|
|
|
|
|
7,096,572 |
|
|
(1)
Includes related party amounts of $1.7 million and $3.8 million for research and development expenses for the six months ended June 30, 2025 and 2026, respectively. See Note 17 — Related Parties.
(2)
Includes related party amounts of $0.6 million and $0.7 million for general and administrative expenses for the six months ended June 30, 2025 and 2026, respectively. See Note 17 — Related Parties.
The accompanying notes are an integral part of these condensed consolidated financial statements
CITY THERAPEUTICS, INC.
CONDENSED CONSOLIDATED STATEMENTS OF CONVERTIBLE PREFERRED STOCK AND STOCKHOLDERS’ DEFICIT
(UNAUDITED, IN THOUSANDS, EXCEPT SHARE AMOUNTS)
| |
|
|
Series Seed
Convertible
Preferred Stock
|
|
|
Series A
Convertible
Preferred
Stock
|
|
|
Series B and B-1
Convertible
Preferred
Stock
|
|
|
|
Common Stock
|
|
|
Additional
Paid-in
Capital
|
|
|
Accumulated
Deficit
|
|
|
Total
Stockholders’
Deficit
|
|
| |
|
|
Shares
|
|
|
Amount
|
|
|
Shares
|
|
|
Amount
|
|
|
Shares
|
|
|
Amount
|
|
|
|
Shares
|
|
|
Amount
|
|
|
Balance as of January 1, 2025
|
|
|
|
|
4,544,990 |
|
|
|
|
$ |
4,475 |
|
|
|
|
|
13,996,318 |
|
|
|
|
$ |
140,039 |
|
|
|
|
|
— |
|
|
|
|
$ |
— |
|
|
|
|
|
|
4,476,828 |
|
|
|
|
$ |
— |
|
|
|
|
$ |
1,174 |
|
|
|
|
$ |
(30,599) |
|
|
|
|
$ |
(29,425) |
|
|
|
Issuance of common stock from exercised options
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
|
|
|
292,075 |
|
|
|
|
|
— |
|
|
|
|
|
184 |
|
|
|
|
|
— |
|
|
|
|
|
184 |
|
|
|
Vesting of restricted common stock
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
|
|
|
423,326 |
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
Stock-based compensation
expense
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
|
|
|
|
|
|
|
|
— |
|
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
|
|
1,497 |
|
|
|
|
|
— |
|
|
|
|
|
1,497 |
|
|
|
Net loss
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
|
|
(24,282) |
|
|
|
|
|
(24,282) |
|
|
|
Balance as of June 30,
2025
|
|
|
|
|
4,544,990 |
|
|
|
|
$ |
4,475 |
|
|
|
|
|
13,996,318 |
|
|
|
|
$ |
140,039 |
|
|
|
|
|
— |
|
|
|
|
$ |
— |
|
|
|
|
|
|
5,192,229 |
|
|
|
|
$ |
— |
|
|
|
|
$ |
2,855 |
|
|
|
|
$ |
(54,881) |
|
|
|
|
$ |
(52,026) |
|
|
| |
|
|
Series Seed
Convertible
Preferred Stock
|
|
|
Series A
Convertible
Preferred
Stock
|
|
|
Series B and B-1
Convertible
Preferred
Stock
|
|
|
|
Common Stock
|
|
|
Additional
Paid-in
Capital
|
|
|
Accumulated
Deficit
|
|
|
Total
Stockholders’
Deficit
|
|
| |
|
|
Shares
|
|
|
Amount
|
|
|
Shares
|
|
|
Amount
|
|
|
Shares
|
|
|
Amount
|
|
|
|
Shares
|
|
|
Amount
|
|
|
Balance as of January 1, 2026
|
|
|
|
|
4,544,990 |
|
|
|
|
$ |
4,475 |
|
|
|
|
|
13,996,318 |
|
|
|
|
$ |
140,039 |
|
|
|
|
|
— |
|
|
|
|
$ |
— |
|
|
|
|
|
|
5,690,551 |
|
|
|
|
$ |
— |
|
|
|
|
$ |
4,449 |
|
|
|
|
$ |
(89,104) |
|
|
|
|
$ |
(84,655) |
|
|
|
Issuance of Series B convertible preferred stock, net of issuance costs of $618
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
|
|
6,801,318 |
|
|
|
|
|
98,882 |
|
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
Conversion of convertible debt to Series B-1 convertible preferred
stock
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
|
|
924,030 |
|
|
|
|
|
12,660 |
|
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
Issuance of common stock from exercised options
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
|
|
|
1,604,104 |
|
|
|
|
|
— |
|
|
|
|
|
1,464 |
|
|
|
|
|
— |
|
|
|
|
|
1,464 |
|
|
|
Vesting of restricted common
stock
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
|
|
|
368,552 |
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
Stock-based compensation
expense
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
|
|
4,917 |
|
|
|
|
|
— |
|
|
|
|
|
4,917 |
|
|
|
Net loss
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
|
|
(53,443) |
|
|
|
|
|
(53,443) |
|
|
|
Balance as of June 30, 2026
|
|
|
|
|
4,544,990 |
|
|
|
|
$ |
4,475 |
|
|
|
|
|
13,996,318 |
|
|
|
|
$ |
140,039 |
|
|
|
|
|
7,725,348 |
|
|
|
|
$ |
111,542 |
|
|
|
|
|
|
7,663,207 |
|
|
|
|
$ |
— |
|
|
|
|
$ |
10,830 |
|
|
|
|
$ |
(142,547) |
|
|
|
|
$ |
(131,717) |
|
|
The accompanying notes are an integral part of these condensed consolidated financial statements
CITY THERAPEUTICS, INC.
CONDENSED CONSOLIDATED STATEMENTS OF CASH FLOWS
(UNAUDITED, IN THOUSANDS)
| |
|
|
Six months ended June 30,
|
|
| |
|
|
2025
|
|
|
2026
|
|
|
Cash flows from operating activities
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
Net loss
|
|
|
|
$ |
(24,282) |
|
|
|
|
$ |
(53,443) |
|
|
|
Adjustments to reconcile net loss to net cash used in operating activities
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
Depreciation
|
|
|
|
|
511 |
|
|
|
|
|
802 |
|
|
|
Stock-based compensation
|
|
|
|
|
1,497 |
|
|
|
|
|
4,917 |
|
|
|
Non-cash interest expense
|
|
|
|
|
646 |
|
|
|
|
|
1,496 |
|
|
|
Gain on lease modification
|
|
|
|
|
(1,024) |
|
|
|
|
|
— |
|
|
|
Change in fair value of convertible notes
|
|
|
|
|
139 |
|
|
|
|
|
2,366 |
|
|
|
Convertible note discount amortization
|
|
|
|
|
7 |
|
|
|
|
|
24 |
|
|
|
Changes in operating assets and liabilities:
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
Accounts receivable
|
|
|
|
|
(313) |
|
|
|
|
|
(3,910) |
|
|
|
Prepaid expenses and other current assets
|
|
|
|
|
(29) |
|
|
|
|
|
1,915 |
|
|
|
Accounts payable, accrued expenses and other current liabilities
|
|
|
|
|
3,440 |
|
|
|
|
|
3,061 |
|
|
|
Deferred revenue
|
|
|
|
|
12,649 |
|
|
|
|
|
(464) |
|
|
|
Other assets
|
|
|
|
|
(11) |
|
|
|
|
|
11 |
|
|
|
Operating lease liabilities
|
|
|
|
|
(328) |
|
|
|
|
|
67 |
|
|
|
Net cash used in operating activities
|
|
|
|
|
(7,098) |
|
|
|
|
|
(43,158) |
|
|
|
Cash flows from investing activities
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
Purchases of property and equipment
|
|
|
|
|
(352) |
|
|
|
|
|
(1,022) |
|
|
|
Net cash used in investing activities
|
|
|
|
|
(352) |
|
|
|
|
|
(1,022) |
|
|
|
Cash flows from financing activities
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
Proceeds from issuance of convertible notes payable.
|
|
|
|
|
30,000 |
|
|
|
|
|
— |
|
|
|
Payment of convertible notes payable issuance costs
|
|
|
|
|
(35) |
|
|
|
|
|
— |
|
|
|
Proceeds from exercise of stock options
|
|
|
|
|
841 |
|
|
|
|
|
2,103 |
|
|
|
Deferred offering costs
|
|
|
|
|
— |
|
|
|
|
|
(224) |
|
|
|
Proceeds from issuance of Series B convertible preferred stock.
|
|
|
|
|
— |
|
|
|
|
|
99,500 |
|
|
|
Payment of Series B convertible preferred stock issuance costs.
|
|
|
|
|
— |
|
|
|
|
|
(387) |
|
|
|
Net cash provided by financing activities
|
|
|
|
|
30,806 |
|
|
|
|
|
100,992 |
|
|
|
Net increase in cash, cash equivalents, and restricted cash
|
|
|
|
|
23,356 |
|
|
|
|
|
56,812 |
|
|
|
Cash, cash equivalents, and restricted cash at beginning of the period
|
|
|
|
|
128,427 |
|
|
|
|
|
115,297 |
|
|
|
Cash, cash equivalents, and restricted cash at end of the period
|
|
|
|
$ |
151,783 |
|
|
|
|
$ |
172,109 |
|
|
|
Supplemental disclosure of noncash investing and financing activities:
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
Property and equipment included in accounts payable and accrued expenses
|
|
|
|
|
48 |
|
|
|
|
|
459 |
|
|
|
Conversion of convertible note and accrued interest into Series B-1 convertible preferred stock
|
|
|
|
|
— |
|
|
|
|
|
12,660 |
|
|
|
Series B convertible preferred stock issuance costs included in accounts payable and accrued expenses
|
|
|
|
|
— |
|
|
|
|
|
231 |
|
|
|
Deferred offering costs included in accounts payable and accrued expenses
|
|
|
|
|
— |
|
|
|
|
|
671 |
|
|
The accompanying notes are an integral part of these condensed consolidated financial statements
CITY THERAPEUTICS, INC.
NOTES TO CONDENSED CONSOLIDATED FINANCIAL STATEMENTS
(UNAUDITED)
1. Nature of Business
Organization
City Therapeutics, Inc. (the Company or City) is a clinical-stage biotechnology company developing the next generation of RNAi therapeutics designed to silence the expression of disease-relevant proteins, with the aim of delivering meaningful and durable benefits for patients in need.
The Company was incorporated in the state of Delaware on July 17, 2023, and has a principal office in Cambridge, Massachusetts.
Liquidity and Capital Resources
Since inception, the Company has devoted substantially all of its efforts to research and development activities, advancing development of the portfolio of programs and platforms, raising capital, developing and maintaining its intellectual property, hiring personnel, and providing general and administrative support for these operations. The Company is subject to risks and uncertainties common to companies in the biotechnology industry, including but not limited to, risks of failure of preclinical studies and clinical trials, development and manufacturing of potential future product candidates, obtaining regulatory approval for potential future product candidates, competition from substitute products, the need to successfully commercialize and gain market acceptance of its potential future product candidates, dependence on key personnel, protection of proprietary technology, compliance with government regulations, development by competitors of technological innovations and the ability to secure additional capital to fund operations.
There can be no assurance that the Company’s research and development efforts will be successfully completed, that adequate protection for the Company’s intellectual property will be obtained, that any products developed will obtain necessary government regulatory approval or that any approved products will be commercially viable. Even if the Company’s product development efforts are successful, it is uncertain when, if ever, the Company will generate significant revenue from product sales. The Company operates in an environment of rapid change in technology and substantial competition from pharmaceutical and biotechnology companies. In addition, the Company is dependent upon the services of its employees and consultants.
The Company has incurred net losses of $24.3 million and $53.4 million during the six months ended June 30, 2025 and 2026, respectively. As of December 31, 2025 and June 30, 2026, the Company had an accumulated deficit of $89.1 million and $142.5 million, respectively. As of June 30, 2026, the Company had cash, cash equivalents, and restricted cash of $172.1 million. To date, the Company’s financing has primarily been through the sale of its Series Seed, Series A and Series B convertible preferred stock, as well as capital received from its collaboration arrangements with Bausch + Lomb Ireland Limited (Bausch + Lomb) and Biogen MA Inc. (Biogen).
The Company has evaluated whether there are conditions and events, considered in the aggregate, that raise substantial doubt about the Company’s ability to continue as a going concern within one year after the date that these condensed consolidated financial statements are issued. As of the issuance date of the condensed consolidated financial statements, the Company expects its cash and cash equivalents will be sufficient to fund its operating expenses and capital expenditure requirements for at least the next twelve months from the date these condensed consolidated financial statements were available to be issued. The future viability of the Company beyond that point is dependent on its ability to raise additional capital to finance its operations. Until such time that the Company can generate significant product revenues, if ever, the Company expects to seek additional funding through private and public equity financings, additional collaborations, strategic alliances and marketing, distribution or licensing arrangements. The Company is seeking to complete an initial public offering (IPO) of its
CITY THERAPEUTICS, INC.
NOTES TO CONDENSED CONSOLIDATED FINANCIAL STATEMENTS (Continued)
(UNAUDITED)
1. Nature of Business (Continued)
common stock. Upon the completion of a qualifying public offering on specified terms, the Company’s outstanding convertible preferred stock will automatically convert into shares of common stock (see Note 10). In the event the Company does not complete an IPO, the Company may not be able to obtain financing on acceptable terms, or at all, and the Company may not be able to enter into additional collaborations or other arrangements. The terms of any financing may adversely affect the holdings or the rights of the Company’s stockholders.
If the Company is unable to obtain sufficient funding, the Company will be forced to delay, scale back or discontinue some or all of its research and development programs, product portfolio expansion efforts or commercialization efforts, which could adversely affect its business prospects. Although management continues to pursue these plans, there is no assurance that the Company will be successful in obtaining sufficient funding on terms acceptable to the Company to fund continuing operations, if at all.
2. Basis of Presentation and Summary of Significant Accounting Policies
The Company’s significant accounting policies are disclosed in Note 2 in the audited consolidated financial statements for the years ended December 31, 2024 and 2025 included elsewhere in this prospectus. Since the date of those financial statements, there have been no changes to the Company’s significant accounting policies, except as noted below.
Unaudited Interim Financial Information
The accompanying interim condensed consolidated financial statements are unaudited. These unaudited interim condensed consolidated financial statements have been prepared on the same basis as the Company’s annual consolidated financial statements and, in the opinion of management, reflect all adjustments that are necessary for the fair statement of the Company’s financial position as of June 30, 2026, and the results of operations and cash flows for the six months ended June 30, 2025 and 2026. The condensed consolidated balance sheet as of December 31, 2025 included herein was derived from the audited consolidated financial statements. The results of operations for the six months ended June 30, 2026 are not necessarily indicative of the results to be expected for the full year ending December 31, 2026, or for any other future annual or interim period.
Deferred Offering Costs
The Company capitalizes certain legal, professional, accounting, and other third-party fees that are directly associated with in-process equity financings as deferred offering costs until such equity financings are consummated. After consummation of the equity financing, these costs are recorded as a reduction in the proceeds from the offering, either as a reduction of the carrying amount of the convertible preferred stock or in stockholders’ deficit as a reduction of additional paid-in capital generated as a result of the offering. Should the in-process equity financing be abandoned, the deferred offering costs are expensed immediately as a charge to operating expenses in the condensed consolidated statement of operations and comprehensive loss. As of December 31, 2025, no deferred offering costs were recorded and as of June 30, 2026, $0.9 million of deferred offering costs were recorded in the condensed consolidated balance sheet.
Common Stock Valuation
Due to the absence of an active public market for the Company’s common stock, the Company utilized methodologies, approaches and assumptions consistent with the American Institute of Certified Public Accountants’ Accounting and Valuation Guide, Valuation of Privately Held Company Equity Securities Issued as Compensation to estimate the fair value of its common stock. In determining the
CITY THERAPEUTICS, INC.
NOTES TO CONDENSED CONSOLIDATED FINANCIAL STATEMENTS (Continued)
(UNAUDITED)
2. Basis of Presentation and Summary of Significant Accounting Policies (Continued)
exercise prices for options granted, the Company has considered the fair value of the common stock as of the grant date. The estimated fair value of the common stock has been determined based upon a variety of factors, including:
•
the prices at which the Company sold convertible preferred stock to new and existing investors and the rights and preferences of the convertible preferred stock relative to our common stock at the time of each grant;
•
the Company’s ability to raise future financings;
•
the progress of the Company’s research and development efforts, including the status of clinical development for our product candidates;
•
the Company’s financial position, including cash on hand, and our historical and forecasted performance and operating results;
•
the Company’s stage of development and business strategy and the material risks related to our business and industry;
•
the achievement of enterprise milestones, including entering into license agreements;
•
any external market conditions affecting the biotechnology industry and trends within the biotechnology industry;
•
the likelihood of achieving a liquidity event for the holders of our convertible preferred stock and holders of our common stock, such as an initial public offering, or a sale of the Company, given prevailing market conditions; and
•
the analysis of initial public offerings and the market performance of similar companies in the biopharmaceutical industry.
Based on the Company’s early stage of development and other relevant factors, options granted prior to June 4, 2026 were valued using an option pricing method (OPM) as an OPM was the most appropriate method for allocating the enterprise value to determine the estimated fair value of the common stock. For options granted after June 4, 2026, the Company determined that a hybrid method that combines both OPM and probability-weighted expected return method (PWERM) was the most appropriate method to determine the estimated fair value of the Company’s common stock.
Significant changes to the key assumptions underlying the factors used could result in different fair values of common stock at each valuation date.
Recently Issued Accounting Standards
In November 2024, the Financial Accounting Standards Board (FASB) issued Accounting Standards Update (ASU) 2024-03, Income Statement — Reporting Comprehensive Income — Expense Disaggregation Disclosures (Subtopic 220-40): Disaggregation of Income Statement Expenses (ASU 2024-03), which is intended to provide more detailed information about specified categories of expenses included in certain expense captions presented on the statement of operations and comprehensive loss. The guidance in ASU 2024-03 is effective for fiscal years beginning after December 15, 2026. Early adoption is permitted. The amendments may be applied either (1) prospectively to financial statements issued for periods after the effective date of ASU 2024-03 or (2) retrospectively to all prior periods presented in the financial statements. The Company is continuing to evaluate the impact that the adoption of ASU 2024-03 may have on its condensed consolidated financial statements and related disclosures for fiscal years beginning after December 15, 2026.
CITY THERAPEUTICS, INC.
NOTES TO CONDENSED CONSOLIDATED FINANCIAL STATEMENTS (Continued)
(UNAUDITED)
2. Basis of Presentation and Summary of Significant Accounting Policies (Continued)
In December 2025, the FASB issued ASU 2025-11, Interim Reporting (Topic 270): Narrow-Scope Improvements (ASU 2025-11) to provide clarifications intended to improve the consistency and usability of interim disclosure requirements. ASU 2025-11 includes a comprehensive listing of required interim disclosures and a new disclosure principle for reporting material events occurring after the most recent annual period. ASU 2025-11 is effective for interim periods within annual reporting periods beginning after December 15, 2027 for public business entities and for interim periods within annual reporting periods beginning after December 15, 2028 for all other entities. Early adoption is permitted. The amendments in this update permit an entity to apply the new guidance using a prospective or retrospective approach. The Company is currently assessing the impact adoption of ASU 2025-11 will have on its condensed consolidated financial statements and related disclosures.
3. Property and Equipment, Net
Property and equipment, net consisted of the following (in thousands):
| |
|
|
December 31,
2025
|
|
|
June 30,
2026
|
|
|
Computer and equipment
|
|
|
|
$ |
198 |
|
|
|
|
$ |
198 |
|
|
|
Furniture and fixtures
|
|
|
|
|
16 |
|
|
|
|
|
16 |
|
|
|
Lab equipment
|
|
|
|
|
5,805 |
|
|
|
|
|
7,287 |
|
|
|
Leasehold improvements
|
|
|
|
|
1,066 |
|
|
|
|
|
1,066 |
|
|
|
Total property and equipment, gross
|
|
|
|
|
7,085 |
|
|
|
|
|
8,567 |
|
|
|
Less: accumulated depreciation
|
|
|
|
|
(1,596) |
|
|
|
|
|
(2,398) |
|
|
|
Property and equipment, net
|
|
|
|
$ |
5,489 |
|
|
|
|
$ |
6,169 |
|
|
Total depreciation expense for the six months ended June 30, 2025 and 2026 was $0.5 million and $0.8 million, respectively.
4. Accrued Expenses and Other Current Liabilities
Accrued expenses and other current liabilities consisted of the following (in thousands):
| |
|
|
December 31,
2025
|
|
|
June 30,
2026
|
|
|
Payroll and employee-related costs
|
|
|
|
$ |
2,480 |
|
|
|
|
$ |
2,566 |
|
|
|
Research and development costs
|
|
|
|
|
2,162 |
|
|
|
|
|
3,586 |
|
|
|
Other
|
|
|
|
|
928 |
|
|
|
|
|
1,847 |
|
|
|
Total accrued expenses and other current liabilities
|
|
|
|
$ |
5,570 |
|
|
|
|
$ |
7,999 |
|
|
5. Convertible Notes
B+L Note
In July 2024, the Company, in connection with the research collaboration and license agreement entered into with Bausch + Lomb (Note 9), issued a convertible note (B+L Note) to Bausch + Lomb for cash proceeds of $10.0 million. The B+L Note was issued at fair value. The B+L Note accrued interest at a rate of 8% per annum. The Company elected the fair value option to account for the B+L Note. The fair value of the B+L Note as of December 31, 2025 was $10.8 million.
CITY THERAPEUTICS, INC.
NOTES TO CONDENSED CONSOLIDATED FINANCIAL STATEMENTS (Continued)
(UNAUDITED)
5. Convertible Notes (Continued)
In May 2026, in connection with the execution of the Company’s Series B Convertible Preferred Stock Purchase Agreement (Series B Agreement), the outstanding principal and accrued interest under the B+L Note were automatically converted into 924,030 shares of Series B-1 convertible preferred stock at a discounted conversion price of $12.4351 per share. This price reflects a 15% discount to the $14.6295 per share purchase price paid by new investors in the Series B financing. The conversion of the B+L Note was accounted for as an extinguishment of debt. The Company recognized the carrying amount of the B+L Note at fair value immediately before conversion into Series B-1 convertible preferred stock (which reflects the fair value of Series B-1 convertible preferred stock issued). As such, no gain or loss was recognized upon conversion.
For the six months ended June 30, 2025 and 2026, the Company recognized $0.4 million and $0.3 million of interest expense related to the B+L Note, respectively. The Company recognized the change in fair value for the B+L Note within total other income, net, in the condensed consolidated statements of operations and comprehensive loss in the amount of a gain of less than $0.1 million and a loss of $0.4 million, inclusive of the change in fair value immediately before conversion, for the six months ended June 30, 2025 and 2026, respectively.
Biogen Note
In May 2025, the Company, in connection with the research collaboration and license agreement entered into with Biogen (Note 9), issued a convertible note (Biogen Note) to Biogen for cash proceeds of $30.0 million. The Biogen Note was issued at fair value. The Biogen Note bears interest at a rate of 8% per annum and matures on July 1, 2027, unless earlier converted into equity securities of the Company. The Company elected the fair value option to account for the Biogen Note. Accordingly, the Company recorded the Biogen Note at fair value upon issuance and at each reporting period until settlement, with changes in fair value recorded in the condensed consolidated statements of operations and comprehensive loss.
The principal and accrued interest under the Biogen Note will automatically convert into shares of the Company’s equity securities upon the earlier of (i) the initial closing of the Company’s next qualified equity financing in which the Company raises at least $100.0 million of cash proceeds or (ii) the closing of the Company’s IPO. In the case of a qualified equity financing, the Biogen Note will convert into shares of preferred stock issued in such financing. In the case of an IPO, the Biogen Note will convert into shares of common stock issued in such offering. If the Biogen Note remains outstanding as of June 30, 2027, the outstanding principal and accrued interest will automatically convert into shares of preferred stock of the Company’s most recently completed equity financing. In each case, the number of shares issued upon conversion will equal the outstanding principal and accrued interest divided by a conversion price equal to 85% of the price paid per share by cash investors in the applicable financing.
For the six months ended June 30, 2025 and 2026, the Company recognized $0.2 million and $1.2 million of interest expense related to the Biogen Note, respectively. The Company recognized a loss on change in fair value for the Biogen Note within total other income, net, in the condensed consolidated statements of operations and comprehensive loss in the amount of $0.2 million and $2.0 million for the six months ended June 30, 2025 and 2026, respectively. The fair value of the Biogen Note as of December 31, 2025 and June 30, 2026 was $31.1 million and $33.1 million, respectively.
CITY THERAPEUTICS, INC.
NOTES TO CONDENSED CONSOLIDATED FINANCIAL STATEMENTS (Continued)
(UNAUDITED)
6. Fair Value of Financial Instruments
The following tables present information about the Company’s financial assets and liabilities that are measured at fair value on a recurring basis and indicate the fair value hierarchy of the valuation techniques the Company utilized to determine such fair value (in thousands):
| |
|
|
December 31, 2025
|
|
| |
|
|
Level 1
|
|
|
Level 2
|
|
|
Level 3
|
|
|
Total
|
|
|
Liabilities
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
Convertible debt, net of discount
|
|
|
|
$ |
— |
|
|
|
|
$ |
— |
|
|
|
|
$ |
41,868 |
|
|
|
|
$ |
41,868 |
|
|
|
Total liabilities
|
|
|
|
$ |
— |
|
|
|
|
$ |
— |
|
|
|
|
$ |
41,868 |
|
|
|
|
$ |
41,868 |
|
|
| |
|
|
June 30, 2026
|
|
| |
|
|
Level 1
|
|
|
Level 2
|
|
|
Level 3
|
|
|
Total
|
|
|
Liabilities
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
Convertible debt, net of discount
|
|
|
|
$ |
— |
|
|
|
|
$ |
— |
|
|
|
|
$ |
33,089 |
|
|
|
|
$ |
33,089 |
|
|
|
Total liabilities
|
|
|
|
$ |
— |
|
|
|
|
$ |
— |
|
|
|
|
$ |
33,089 |
|
|
|
|
$ |
33,089 |
|
|
There have been no transfers between fair value levels during the six months ended June 30, 2025 and 2026.
Convertible Notes
As further described in Note 5, in July 2024, the Company issued the B+L Note in connection with a research collaboration and license agreement with Bausch + Lomb. In May 2025, the Company issued the Biogen Note in connection with a research collaboration and license agreement with Biogen. The Company elected the fair value option for each of the convertible note issuances. The fair value of each of the convertible notes was estimated using a scenario-based analysis that estimates the fair value based on the probability-weighted present value of expected future investment returns, considering each of the possible outcomes available to the noteholder. The Biogen Note remains outstanding as of June 30, 2026.
The B+L Note, as described in Note 5, was converted into Series B-1 convertible preferred stock in May 2026. The final fair value of the B+L Note was determined using an OPM backsolve to the Series B convertible preferred stock, considering the differences in liquidation preference between the Series B and Series B-1 convertible preferred stock.
The following tables summarize information about the significant unobservable inputs used in the fair value measurements for the B+L Note and the Biogen Note:
CITY THERAPEUTICS, INC.
NOTES TO CONDENSED CONSOLIDATED FINANCIAL STATEMENTS (Continued)
(UNAUDITED)
6. Fair Value of Financial Instruments (Continued)
| |
|
|
December 31, 2025
|
|
| |
|
|
Fair value as
of initial
valuation
date
(in thousands)
|
|
|
Fair value as
of period end
(in thousands)
|
|
|
Expected
settlement
options (and
relative
weighting)
|
|
|
Key
unobservable
inputs
|
|
|
Range
|
|
|
B + L Note
|
|
|
|
$ |
10,000 |
|
|
|
|
$ |
10,805 |
|
|
|
Equity
financing (95)%
|
|
|
Estimated
time to
settlement:
Discount rate:
|
|
|
0.42 years
18.17%
|
|
|
Biogen Note
|
|
|
|
$ |
30,000 |
|
|
|
|
$ |
31,063 |
|
|
|
Equity
financing (95)%
|
|
|
Estimated
time to
settlement:
Discount rate:
|
|
|
1.25 years
14.39%
|
|
| |
|
|
June 30, 2026
|
|
| |
|
|
Fair value as
of initial
valuation
date
(in thousands)
|
|
|
Fair value as
of period end
(in thousands)
|
|
|
Expected
settlement
options (and
relative
weighting)
|
|
|
Key
unobservable
inputs
|
|
|
Range
|
|
|
Biogen Note
|
|
|
|
$ |
30,000 |
|
|
|
|
$ |
33,089 |
|
|
|
Equity financing (95)%
|
|
|
Estimated
time to
settlement:
Discount rate:
|
|
|
0.33 years
14.39%
|
|
The following table provides a reconciliation of all liabilities measured at fair value using Level 3 significant unobservable inputs (in thousands):
| |
|
|
Convertible
Notes
|
|
|
Balance at January 1, 2025
|
|
|
|
$ |
10,276 |
|
|
|
Issuance of convertible note, net on May 23, 2025
|
|
|
|
|
29,965 |
|
|
|
Amortization of issuance costs.
|
|
|
|
|
21 |
|
|
|
Change in fair value of convertible notes
|
|
|
|
|
1,606 |
|
|
|
Balance at December 31, 2025
|
|
|
|
$ |
41,868 |
|
|
|
Amortization of issuance costs.
|
|
|
|
|
24 |
|
|
|
Change in fair value of convertible notes
|
|
|
|
|
2,366 |
|
|
|
Non-cash conversion of B+L note to Series B-1 convertible preferred stock
|
|
|
|
|
(11,169) |
|
|
|
Balance at June 30, 2026
|
|
|
|
$ |
33,089 |
|
|
During the six months ended June 30, 2025 and 2026, there were no transfers between levels.
CITY THERAPEUTICS, INC.
NOTES TO CONDENSED CONSOLIDATED FINANCIAL STATEMENTS (Continued)
(UNAUDITED)
7. Leases
In June 2025, the Company entered into the First Amendment to Lease (the Amendment), which expanded the leased premises to include the entire second floor and a portion of the fourth floor, provided for the surrender of the original first floor and penthouse mechanical premises and extended the lease term through September 30, 2031. Following the Amendment, the Company’s leased premises consist of approximately 51,726 rentable square feet. The Company recorded a gain on lease modification of $1.0 million in other income, net in our condensed consolidated statement of operations and comprehensive loss.
There were no material lease modifications, impairments, or new lease arrangements during the six months ended June 30, 2026.
The elements of operating lease expense were as follows (in thousands):
| |
|
|
Six months ended
June 30,
|
|
| |
|
|
2025
|
|
|
2026
|
|
|
Lease cost
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
Operating lease cost
|
|
|
|
$ |
1,471 |
|
|
|
|
$ |
3,140 |
|
|
|
Variable lease cost
|
|
|
|
|
701 |
|
|
|
|
|
1,237 |
|
|
|
Total lease cost
|
|
|
|
$ |
2,172 |
|
|
|
|
$ |
4,377 |
|
|
8. Commitments and Contingencies
Litigation
Liabilities for loss contingencies arising from claims, assessments, litigation, fines, penalties, and other sources are recorded when it is probable that a liability has been incurred and the amount can be reasonably estimated. From time to time, the Company may become involved in legal proceedings arising in the ordinary course of business. As of December 31, 2025 and June 30, 2026, the Company had no matters for which a loss was considered probable or reasonably possible that would require accrual or disclosure under ASC 450, Contingencies.
Indemnification
In the ordinary course of business, the Company may provide indemnification of varying scope and terms to vendors, lessors, business partners and other parties with respect to certain matters including, but not limited to, losses arising out of breach of such agreements or from intellectual property infringement claims made by third parties. In addition, the Company has entered into, or intends to enter into, indemnification agreements with all members of the board of directors (Board) and officers that will require the Company, among other things, to indemnify them against certain liabilities that may arise by reason of their status or service as directors.
The maximum potential amount of future payments the Company could be required to make under these indemnification agreements is not determinable. To date, the Company has not incurred any material costs as a result of such indemnifications. The Company is not aware of any claims under indemnification arrangements that could have a material effect on its financial position, results of operations or cash flows, and it has not accrued any liabilities related to such obligations in its condensed consolidated financial statements as of December 31, 2025 and June 30, 2026.
CITY THERAPEUTICS, INC.
NOTES TO CONDENSED CONSOLIDATED FINANCIAL STATEMENTS (Continued)
(UNAUDITED)
9. Significant Agreements
Ohio State Innovation Foundation License Agreement
On October 30, 2023, the Company entered into an exclusive license agreement (as amended, the OSIF License Agreement) with the Ohio State Innovation Foundation (OSIF), pursuant to which the Company has been granted an exclusive, royalty-bearing license to certain patent rights to make, have made, use, sell, and import licensed products and technology disclosures to the licensed intellectual property.
Pursuant to the OSIF License Agreement, the Company is required to make milestone payments of up to $4.2 million upon achievement of certain clinical development, regulatory approval, and commercial sales milestones for each licensed product. The Company is also required to pay royalties to OSIF in the low-single digit percentage range based on net sales of any licensed products and processes, along with a specified minimum annual royalty amount for each licensed product, subject to customary reductions and offsets. In addition, the Company is responsible for certain patent costs incurred after the effective date and annual maintenance fees. The OSIF License Agreement also requires the Company to pay OSIF tiered percentages of certain non-royalty sublicense consideration, ranging from a low-twenties to mid-twenties percentage at earlier stages of development and a low-tens to mid-teens percentage at later stages of development, with the applicable percentage depending on the stage of development at which the sublicense is granted. In connection with the Biogen Research Collaboration and License Agreement (Note 9), the Company accrued $0.3 million in sublicensing fees payable to OSIF during the six months ended June 30, 2025 and no payments were made or accrued for the six months ended June 30, 2026. No milestone payments or net sales-based royalties had become due to OSIF through June 30, 2026. No repurchase rights of common stock issued to OSIF had become payable as of June 30, 2026. No acquired in-process research and development expense was recognized in connection with the OSIF License Agreement during the six months ended June 30, 2025 and 2026, respectively.
Bausch + Lomb Research Collaboration and License Agreement
On July 8, 2024, the Company entered into a research collaboration and license agreement (B+L Research Collaboration and License Agreement) with Bausch + Lomb, pursuant to which the Company is performing research activities to identify development candidates for ocular diseases and granted Bausch + Lomb an option to obtain an exclusive worldwide license to certain Company intellectual property.
Pursuant to the B+L Research Collaboration and License Agreement, the Company will perform research and deliver to Bausch + Lomb a development candidate for which Bausch + Lomb may exercise their option to license. If Bausch + Lomb exercises its option, it will engage in the development, manufacturing and commercialization of licensed molecules and licensed products from the development candidate. Under the B+L Research Collaboration and License Agreement, the Company received a non- refundable upfront payment of $5.0 million from Bausch + Lomb. Over the course of the research plan, Bausch + Lomb is required to reimburse the Company for amounts incurred for full-time employees and other related costs and materials directly associated with the research plan. Additionally, the Company is eligible to receive a certain development candidate selection fee and certain other development, regulatory, commercial and sales milestones, collectively up to an aggregate of $485.0 million, along with tiered royalties ranging in the mid-single digits to low teens on net sales. If the agreement terminates and the Company later successfully commercializes a product candidate using the licensed intellectual property, the Company will owe a low-single digit royalty payment to Bausch + Lomb on future net sales for a fixed royalty term. As of June 30, 2026, no such milestone payments or royalties had become due to the Company.
CITY THERAPEUTICS, INC.
NOTES TO CONDENSED CONSOLIDATED FINANCIAL STATEMENTS (Continued)
(UNAUDITED)
9. Significant Agreements (Continued)
The Company accounts for the agreement as a single performance obligation satisfied over time using a cost-based input method. As of June 30, 2026, the total transaction price is $15.0 million.
Biogen Research Collaboration and License Agreement
On May 23, 2025, the Company entered into a research collaboration and license agreement (Biogen Research Collaboration and License Agreement) with Biogen, pursuant to which (i) the Company and Biogen shall collaborate to undertake a research plan to discover certain RNAi triggers that inhibit the expression of a target protein and conjugates of such RNAi triggers and (ii) the Company granted Biogen an exclusive, worldwide, royalty-bearing license to certain Company intellectual property to develop, manufacture and commercialize certain licensed products in the field in the territory.
Pursuant to the Biogen Research Collaboration and License Agreement, the Company will perform research activities and deliver to Biogen data packages and technology transfers to enable Biogen to select and advance a development candidate, for which Biogen may exercise its rights under the license. If Biogen selects a development candidate, it will control the development, manufacturing and commercialization of licensed compounds and licensed products. Each party is responsible for its own research costs under the agreement. Biogen has the option of selecting one additional target during the option exercise period, subject to execution of a separate agreement and payment of a specified option exercise fee. The Company received a $16.0 million upfront payment and may receive an additional upfront payment upon the satisfaction of certain conditions relating to the Company’s upstream third-party intellectual property agreement, as well as development, commercial, and sales milestone payments of up to approximately $1.0 billion, and tiered royalties from the high-single digits to low-teens on net sales. As of June 30, 2026, no such milestones or royalties had become due to the Company.
The Company accounts for the agreement as a single performance obligation satisfied over time using a cost-based input method. At contract inception, the transaction price was determined to be $16.0 million, which represents the nonrefundable upfront payment under the Biogen Research Collaboration and License Agreement. The $4.0 million additional upfront payment, research milestones, development milestones, sales milestones, and royalties are variable consideration that is fully constrained at inception and therefore excluded from the initial transaction price. During the year ended December 31, 2025, one research milestone of $2.5 million became probable and was included in the transaction price, increasing the total transaction price to $18.5 million as of December 31, 2025.
During the period ended June 30, 2026, the Company and Biogen agreed to increase the research milestone from $2.5 million to $4.0 million. The Company concluded this modification did not result in a distinct new promise or good, resulting in the Company maintaining its single performance obligation under the Biogen Research Collaboration and License Agreement. This increase was accounted for as a modification and the transaction price was revised to $20.0 million, resulting in a cumulative catch-up adjustment to revenue in the six months ended June 30, 2026.
CITY THERAPEUTICS, INC.
NOTES TO CONDENSED CONSOLIDATED FINANCIAL STATEMENTS (Continued)
(UNAUDITED)
9. Significant Agreements (Continued)
Amounts recognized under the B+L Research Collaboration and License Agreement and Biogen Research Collaboration and License Agreement were as follows (in thousands):
| |
|
|
Six months
ended
June 30,
|
|
| |
|
|
2025
|
|
|
2026
|
|
|
Revenue recognized from B+L Research Collaboration and License Agreement (Ocular)
|
|
|
|
$ |
3,567 |
|
|
|
|
$ |
2,845 |
|
|
|
Revenue recognized from Biogen Research Collaboration and License Agreement (CNS)
|
|
|
|
|
2,011 |
|
|
|
|
|
3,513 |
|
|
|
Total Revenue
|
|
|
|
$ |
5,578 |
|
|
|
|
$ |
6,358 |
|
|
| |
|
|
December 31,
2025
|
|
|
June 30,
2026
|
|
|
Amounts receivable under B+L Research Collaboration and License Agreement
|
|
|
|
$ |
1,209 |
|
|
|
|
$ |
5,119 |
|
|
|
Deferred revenue, current portion
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
Deferred revenue – Bausch + Lomb, current portion
|
|
|
|
|
1,693 |
|
|
|
|
|
743 |
|
|
|
Deferred revenue – Biogen, current portion
|
|
|
|
|
9,562 |
|
|
|
|
|
10,048 |
|
|
|
Total Deferred Revenue, current portion
|
|
|
|
$ |
11,255 |
|
|
|
|
$ |
10,791 |
|
|
|
Deferred revenue, non-current portion
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
Deferred revenue – Bausch + Lomb, noncurrent portion
|
|
|
|
$ |
— |
|
|
|
|
$ |
— |
|
|
|
Deferred revenue – Biogen, noncurrent portion
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
Total Deferred Revenue, non-current portion
|
|
|
|
$ |
— |
|
|
|
|
$ |
— |
|
|
The accounts receivable recorded as of December 31, 2025 and June 30, 2026 related to the reimbursable research and development efforts by the Company which were invoiced as incurred.
The following table summarizes the changes in deferred revenue related to the upfront non-refundable payments received as part of the B+L Research Collaboration and License Agreement and Biogen Research Collaboration and License Agreement (in thousands):
| |
|
|
Six months
ended
June 30,
|
|
| |
|
|
2025
|
|
|
2026
|
|
|
Beginning balance
|
|
|
|
$ |
3,956 |
|
|
|
|
$ |
11,255 |
|
|
|
Additions
|
|
|
|
|
16,000 |
|
|
|
|
|
4,000 |
|
|
|
Deductions
|
|
|
|
|
(3,351) |
|
|
|
|
|
(4,464) |
|
|
|
Ending balance
|
|
|
|
$ |
16,605 |
|
|
|
|
$ |
10,791 |
|
|
CITY THERAPEUTICS, INC.
NOTES TO CONDENSED CONSOLIDATED FINANCIAL STATEMENTS (Continued)
(UNAUDITED)
9. Significant Agreements (Continued)
The Company recognizes the following revenues from amounts included in deferred revenue as of the beginning of the period (in thousands):
| |
|
|
Six months
ended
June 30,
|
|
| |
|
|
2025
|
|
|
2026
|
|
|
Revenue from amounts included in deferred revenue as of the beginning of the period
|
|
|
|
$ |
1,341 |
|
|
|
|
$ |
3,344 |
|
|
10. Convertible Preferred Stock
Series Seed Convertible Preferred Stock
Beginning in September 2023 and in multiple closings through March 2024, the Company issued shares of its Series Seed convertible preferred stock pursuant to a Series Seed Convertible Preferred Stock Purchase Agreement (Series Seed Agreement) with certain investors. The Company issued a total of 4,544,990 shares of Series Seed convertible preferred stock at a purchase price of $1.00 per share for gross proceeds of $4.5 million across multiple closings. The Company incurred $0.1 million in issuance costs associated with the Series Seed convertible preferred stock financing.
Series A Convertible Preferred Stock
Beginning in April 2024 and closing in multiple closings through December 2024, the Company issued shares of its Series A convertible preferred stock pursuant to the Series A Convertible Preferred Stock Purchase Agreement (Series A Agreement) with certain investors. The Company issued an aggregate of 13,996,318 shares of Series A convertible preferred stock. Of the 13,996,318 shares issued, 13,435,354 shares were issued at a price per share of $10.0295 for aggregate gross proceeds of $134.7 million. Additionally, the Company’s outstanding convertible notes and related accrued interest, at the time of the Series A Agreement, totaling $5.1 million, converted into 560,964 shares of Series A convertible preferred stock at a price per share of $9.02655.
Series B and B-1 Convertible Preferred Stock
Beginning in May 2026 and in one additional closing in June 2026, the Company issued shares of its Series B and B-1 convertible preferred stock pursuant to the Series B Agreement with certain investors. The Company issued an aggregate of 7,725,348 shares of Series B and B-1 convertible preferred stock. Of the 7,725,348 shares issued, 6,801,318 shares of Series B convertible preferred stock were issued at a price per share of $14.6295 for aggregate gross proceeds of $99.5 million. Additionally, the Company’s outstanding B+L Note and related accrued interest, at the time of the Series B Agreement, totaling $11.5 million, converted into 924,030 shares of Series B-1 convertible preferred stock at a price per share of $12.4351 with an aggregate fair value of $12.7 million (or $13.70 per share of Series B-1 convertible preferred stock). See Note 5.
CITY THERAPEUTICS, INC.
NOTES TO CONDENSED CONSOLIDATED FINANCIAL STATEMENTS (Continued)
(UNAUDITED)
10. Convertible Preferred Stock (Continued)
The Company’s convertible preferred stock is summarized below (in thousands, except share amounts):
| |
|
|
December 31, 2025
|
|
| |
|
|
Total Shares
Authorized
|
|
|
Total Shares
Issued and
Outstanding
|
|
|
Common
Shares
Issuable upon
Conversion
|
|
|
Carrying
Value
|
|
|
Liquidation
Preference
|
|
|
Series Seed convertible preferred stock
|
|
|
|
|
4,544,990 |
|
|
|
|
|
4,544,990 |
|
|
|
|
|
4,991,760 |
|
|
|
|
$ |
4,475 |
|
|
|
|
$ |
4,545 |
|
|
|
Series A convertible preferred stock
|
|
|
|
|
14,295,436 |
|
|
|
|
|
13,996,318 |
|
|
|
|
|
15,372,145 |
|
|
|
|
|
140,039 |
|
|
|
|
|
140,376 |
|
|
|
Total
|
|
|
|
|
18,840,426 |
|
|
|
|
|
18,541,308 |
|
|
|
|
|
20,363,905 |
|
|
|
|
$ |
144,514 |
|
|
|
|
$ |
144,921 |
|
|
| |
|
|
June 30, 2026
|
|
| |
|
|
Total Shares
Authorized
|
|
|
Total Shares
Issued and
Outstanding
|
|
|
Common
Shares
Issuable upon
Conversion
|
|
|
Carrying
Value
|
|
|
Liquidation
Preference
|
|
|
Series Seed convertible preferred stock
|
|
|
|
|
4,544,990 |
|
|
|
|
|
4,544,990 |
|
|
|
|
|
4,991,760 |
|
|
|
|
$ |
4,475 |
|
|
|
|
$ |
4,545 |
|
|
|
Series A convertible preferred stock
|
|
|
|
|
13,996,318 |
|
|
|
|
|
13,996,318 |
|
|
|
|
|
15,372,145 |
|
|
|
|
|
140,039 |
|
|
|
|
|
140,376 |
|
|
|
Series B convertible preferred stock
|
|
|
|
|
6,801,318 |
|
|
|
|
|
6,801,318 |
|
|
|
|
|
7,469,874 |
|
|
|
|
|
98,882 |
|
|
|
|
|
99,500 |
|
|
|
Series B-1 convertible preferred stock
|
|
|
|
|
924,030 |
|
|
|
|
|
924,030 |
|
|
|
|
|
1,014,862 |
|
|
|
|
|
12,660 |
|
|
|
|
|
11,490 |
|
|
|
Total
|
|
|
|
|
26,266,656 |
|
|
|
|
|
26,266,656 |
|
|
|
|
|
28,848,641 |
|
|
|
|
$ |
256,056 |
|
|
|
|
$ |
255,911 |
|
|
The holders of the Series Seed, A, B, and B-1 convertible preferred stock have the following rights, preferences and privileges:
Conversion
All series of convertible preferred stock are convertible at any time at the option of the holder and mandatorily convertible upon a) a firm-commitment underwritten qualified initial public offering resulting in at least $75.0 million of gross proceeds to the Company (Qualified IPO) or b) the vote or written consent of holders of at least 56% of the outstanding shares of convertible preferred stock (Requisite Holders) into common stock. The conversion ratio of each series of the convertible preferred stock is determined by dividing the original issuance price of each series by the applicable conversion price of each series. The Series Seed, A, B, and B-1 convertible preferred stock conversion price initially is equal to $0.9105, $9.1319, $13.3202, and $11.3222 per share, respectively, each subject to appropriate adjustment in the event of any stock dividend, stock split, combination or other similar recapitalization and other adjustments, including adjustment if common stock is issued for less than the original issue price of each series of convertible preferred stock (subject to certain exceptions, including shares of common stock in a Qualified IPO).
CITY THERAPEUTICS, INC.
NOTES TO CONDENSED CONSOLIDATED FINANCIAL STATEMENTS (Continued)
(UNAUDITED)
10. Convertible Preferred Stock (Continued)
Voting
The holder of each share of Series Seed, A, B, and B-1 convertible preferred stock is entitled to one vote for each share of common stock into which such shares of Series Seed, A, B, and B-1 convertible preferred stock could then be converted and votes together with the holders of common stock as a single class, on an as-converted to common stock basis.
Redemption
The holders of Series Seed, A, B, and B-1 convertible preferred stock do not have any redemption rights, except for the contingent redemption upon the occurrence of a Deemed Liquidation Event (as defined below).
Liquidation
In the event of any voluntary or involuntary liquidation, dissolution or winding up of the Company, or upon the occurrence of a Deemed Liquidation Event (as defined below), the holders of the Series Seed, A, B, and B-1 convertible preferred stock are entitled to receive an amount per share equal to the greater of (i) the respective original issuance price, plus any dividends declared but unpaid or (ii) the amount payable with respect to such share if it was converted to common stock immediately prior to such liquidation, dissolution, winding up or Deemed Liquidation Event (as defined below). Liquidation payments to the holders of the Series Seed, A, B, and B-1 convertible preferred stock have priority and are made in preference to any payments to the holders of common stock. After full payment of the liquidation preference to the holders of the Series Seed, A, B, and B-1 convertible preferred stock, the remaining assets, if any, will be distributed to the holders of shares of common stock, pro rata based on the number of shares held by each holder. Unless the Requisite Holders, elect otherwise, a “Deemed Liquidation Event” shall include a merger, consolidation, or share exchange (other than one in which stockholders of the Company own a majority by voting power of the outstanding shares of the surviving or acquiring corporation) or a sale, lease, transfer, exclusive license or other disposition of all or substantially all of the assets of the Company taken as a whole or the sale or disposition of one or more subsidiaries if substantially all of the assets of the Company and its subsidiaries are held by such subsidiary or subsidiaries.
Dividends
The Company shall not declare, pay or set aside any dividends on shares of any class of capital stock of the Company unless the holders of the Series Seed, A, B, and B-1 convertible preferred stock shall first receive, or simultaneously receive, a noncumulative dividend on each outstanding share of the Series Seed, A, B, and B-1 convertible preferred stock equal to 8% of the original issue prices. No such dividends have been declared or paid through June 30, 2026.
11. Common Stock
During the six months ended June 30, 2026, the Company increased the number of authorized shares of common stock from 30,000,000 to 42,350,000 shares of $0.0001 par value common stock.
Features of the Common Stock
The common stock has a par value of $0.0001, and the holders of common stock are entitled to one vote for each share of common stock held at all meetings of stockholders and written actions in lieu of meetings provided. All dividends shall be declared and paid pro rata according to the number of shares
CITY THERAPEUTICS, INC.
NOTES TO CONDENSED CONSOLIDATED FINANCIAL STATEMENTS (Continued)
(UNAUDITED)
11. Common Stock (Continued)
held by each stockholder. In the event of a liquidation, dissolution, or winding up of the Company, the common stock ranks behind the preferred stock.
Common stock reserved for future issuance consists of the following:
| |
|
|
December 31,
2025
|
|
|
June 30,
2026
|
|
|
Series Seed convertible preferred stock
|
|
|
|
|
4,991,760 |
|
|
|
|
|
4,991,760 |
|
|
|
Series A convertible preferred stock
|
|
|
|
|
15,372,145 |
|
|
|
|
|
15,372,145 |
|
|
|
Series B convertible preferred stock
|
|
|
|
|
— |
|
|
|
|
|
7,469,874 |
|
|
|
Series B-1 convertible preferred stock
|
|
|
|
|
— |
|
|
|
|
|
1,014,862 |
|
|
|
Unvested restricted stock
|
|
|
|
|
2,081,638 |
|
|
|
|
|
1,964,421 |
|
|
|
Stock options issued and outstanding
|
|
|
|
|
4,768,315 |
|
|
|
|
|
5,404,513 |
|
|
|
Shares available for issuance under the Plan
|
|
|
|
|
624,683 |
|
|
|
|
|
1,482,874 |
|
|
|
Total
|
|
|
|
|
27,838,541 |
|
|
|
|
|
37,700,449 |
|
|
12. Stock-based Compensation
2023 Stock Plan
The 2023 Plan was originally adopted with 1,136,740 shares of common stock reserved for issuance. The Board subsequently increased the number of shares reserved under the 2023 Plan at various times during 2024, 2025, and 2026. As of June 30, 2026, the Company had reserved 10,017,776 shares of common stock for issuance under the 2023 Plan. There are no additional shares of common stock available for future issuance outside the common stock reserved under the 2023 Plan.
The stock options granted to employees follow various vesting periods ranging from immediate to four-year vesting and have a term of up to ten years from the date of grant.
Stock Option Valuation
The Company estimates the fair value of stock options using the Black-Scholes option-pricing model. This model incorporates various assumptions, including the expected volatility, expected term, and interest rates.
The following table presents, on a weighted-average basis, the assumptions used in the Black-Scholes option-pricing model to determine the fair value of stock options granted during the six months ended June 30, 2025 and 2026:
| |
|
|
Six months ended June 30,
|
|
| |
|
|
2025
|
|
|
2026
|
|
|
Risk-free interest rate
|
|
|
4.12% – 4.22%
|
|
|
3.64% – 4.26%
|
|
|
Dividend yield
|
|
|
0%
|
|
|
0%
|
|
|
Expected term of options (years) range
|
|
|
5.27 – 6.08
|
|
|
5.23 – 6.08
|
|
|
Volatility rate range
|
|
|
87.73% – 111.06%
|
|
|
100.27% – 108.11%
|
|
The weighted-average grant-date fair value of stock options granted during the six months ended June 30, 2025 and 2026 was $2.07 and $2.97 per share, respectively.
CITY THERAPEUTICS, INC.
NOTES TO CONDENSED CONSOLIDATED FINANCIAL STATEMENTS (Continued)
(UNAUDITED)
12. Stock-based Compensation (Continued)
Time-Based Stock Options
The following table summarizes the Company’s time-based stock option activity for the six months ended June 30, 2026:
| |
|
|
Number of
Outstanding
Options
|
|
|
Weighted
Average
Exercise
Price
|
|
|
Weighted
Average
Remaining
Contractual
Term
(in years)
|
|
|
Aggregate
Intrinsic
Value
|
|
|
Balance at December 31, 2025
|
|
|
|
|
3,367,868 |
|
|
|
|
$ |
1.13 |
|
|
|
|
|
8.98 |
|
|
|
|
$ |
2,383 |
|
|
|
Granted
|
|
|
|
|
1,263,462 |
|
|
|
|
|
3.93 |
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
Exercised
|
|
|
|
|
(1,220,557) |
|
|
|
|
|
1.16 |
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
Cancelled
|
|
|
|
|
— |
|
|
|
|
|
— |
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
Forfeited
|
|
|
|
|
(35,754) |
|
|
|
|
|
1.65 |
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
Balance at June 30, 2026
|
|
|
|
|
3,375,019 |
|
|
|
|
$ |
2.16 |
|
|
|
|
|
8.94 |
|
|
|
|
$ |
19,341 |
|
|
|
Vested and expected to vest as of June 30, 2026
|
|
|
|
|
3,375,019 |
|
|
|
|
$ |
2.16 |
|
|
|
|
|
8.94 |
|
|
|
|
$ |
19,341 |
|
|
|
Vested and exercisable at June 30, 2026
|
|
|
|
|
753,612 |
|
|
|
|
$ |
0.94 |
|
|
|
|
|
8.23 |
|
|
|
|
$ |
5,243 |
|
|
As of June 30, 2026, there was $7.9 million of total unrecognized compensation cost related to unvested time-based stock options, and the Company expects to recognize such amount over a remaining weighted-average period of 3.3 years.
Performance-Based Stock Options
The Company has granted stock options to certain employees to purchase shares of common stock that contain performance-based vesting criteria related to corporate or scientific milestones. The fair value of each option grant with performance-based vesting was estimated on the date of grant.
Recognition of stock-based compensation expense associated with these performance-based stock options commences when the performance condition is considered probable of achievement, using management’s best estimates, which consider the inherent risk and uncertainty regarding the future outcomes of the milestones. In January 2026, the Company determined that the achievement of a scientific milestone was probable, triggering the vesting condition on certain performance-based stock options. As a result, the Company recognized $3.4 million of stock-based compensation expense related to the vesting of 1,071,044 stock options for the six months ended June 30, 2026.
CITY THERAPEUTICS, INC.
NOTES TO CONDENSED CONSOLIDATED FINANCIAL STATEMENTS (Continued)
(UNAUDITED)
12. Stock-based Compensation (Continued)
The following table summarizes the Company’s performance-based stock option activity for the six months ended June 30, 2026:
| |
|
|
Number of
Outstanding
Options
|
|
|
Weighted
Average
Exercise
Price
|
|
|
Weighted
Average
Remaining
Contractual
Term
(in years)
|
|
|
Aggregate
Intrinsic
Value
|
|
|
Balance at December 31, 2025
|
|
|
|
|
1,400,447 |
|
|
|
|
$ |
1.13 |
|
|
|
|
|
9.02 |
|
|
|
|
$ |
1,000 |
|
|
|
Granted
|
|
|
|
|
1,265,324 |
|
|
|
|
|
1.96 |
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
Exercised
|
|
|
|
|
(634,895) |
|
|
|
|
|
1.12 |
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
Forfeited
|
|
|
|
|
(1,382) |
|
|
|
|
|
1.23 |
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
Balance at June 30, 2026
|
|
|
|
|
2,029,494 |
|
|
|
|
$ |
1.65 |
|
|
|
|
|
9.19 |
|
|
|
|
$ |
12,682 |
|
|
|
Vested and exercisable at June 30, 2026
|
|
|
|
|
210,137 |
|
|
|
|
$ |
0.92 |
|
|
|
|
|
8.24 |
|
|
|
|
$ |
1,466 |
|
|
As of June 30, 2026, there was $3.5 million of total unrecognized compensation cost related to unvested performance-based stock options, and the Company expects to recognize such amount when the achievement of the underlying performance conditions becomes probable. Because vesting of the performance-based options is not reasonably assured, the Company does not consider these options to be expected to vest.
Restricted Common Stock Awards
The Company awarded restricted common stock to employees and non-employees under its 2023 Plan and may continue to award restricted common stock to employees and non-employees under the 2023 Plan. Additionally, employees who exercise their option awards prior to the vesting of that award hold common stock that is subject to further time- or performance-based vesting conditions and therefore classified as restricted stock for accounting purposes. The fair value of each share of restricted common stock is based on the estimated fair value of the Company’s common stock on the date of grant.
Time-Based Restricted Common Stock Awards
Restricted common stock that has time-based vesting conditions vest over a four-year period, subject to the employee’s continued employment with, or service to, the Company on each vesting date. Compensation expense is recognized on a straight-line basis over the vesting period.
The following table summarizes the Company’s time-based restricted common stock award activity for the six months ended June 30, 2026:
| |
|
|
Number
of
Shares
|
|
|
Weighted
Average
Grant Date
Fair Value
|
|
|
Unvested restricted common stock at December 31, 2025
|
|
|
|
|
1,703,539 |
|
|
|
|
$ |
1.02 |
|
|
|
Issued
|
|
|
|
|
755,428 |
|
|
|
|
|
2.38 |
|
|
|
Vested
|
|
|
|
|
(646,633) |
|
|
|
|
|
1.15 |
|
|
|
Unvested restricted common stock at June 30, 2026
|
|
|
|
|
1,812,334 |
|
|
|
|
$ |
1.54 |
|
|
CITY THERAPEUTICS, INC.
NOTES TO CONDENSED CONSOLIDATED FINANCIAL STATEMENTS (Continued)
(UNAUDITED)
12. Stock-based Compensation (Continued)
As of June 30, 2026, there was $3.7 million of total unrecognized compensation cost related to time-based restricted common stock, and the Company expects to recognize such amount over a remaining weighted-average period of 2.3 years.
Performance-Based Restricted Common Stock Awards
Restricted common stock that have performance-based vesting conditions vest when certain milestones are achieved and approved by the Board, subject to the employee’s continued employment with, or service to, the Company on each vesting date. Compensation expense is recognized upon the achievement of the certain milestones.
The following table summarizes the Company’s performance-based restricted common stock award activity for the six months ended June 30, 2026:
| |
|
|
Number of
Shares
|
|
|
Weighted
Average
Grant Date
Fair Value
|
|
|
Unvested restricted common stock at December 31, 2025
|
|
|
|
|
378,099 |
|
|
|
|
$ |
3.65 |
|
|
|
Issued
|
|
|
|
|
150,869 |
|
|
|
|
|
4.94 |
|
|
|
Vested
|
|
|
|
|
(376,881) |
|
|
|
|
|
3.65 |
|
|
|
Unvested restricted common stock at June 30, 2026
|
|
|
|
|
152,087 |
|
|
|
|
$ |
4.91 |
|
|
As of June 30, 2026, there was $0.2 million of total unrecognized compensation cost related to performance-based restricted common stock, and the Company expects to recognize such amount when the achievement of the underlying performance conditions become probable.
Stock-Based Compensation
The following table below summarizes the classification of the Company’s stock-based compensation expense related to stock options and restricted common stock awards in the condensed consolidated statements of operations and comprehensive loss (in thousands):
| |
|
|
Six months ended
June 30,
|
|
| |
|
|
2025
|
|
|
2026
|
|
|
Research and development
|
|
|
|
$ |
442 |
|
|
|
|
$ |
1,316 |
|
|
|
General and administrative
|
|
|
|
|
1,055 |
|
|
|
|
|
3,601 |
|
|
|
Total stock-based compensation expense
|
|
|
|
$ |
1,497 |
|
|
|
|
$ |
4,917 |
|
|
13. 401(k) Plan
The Company has a defined-contribution savings plan under Section 401(k) of the IRC (401(k) Plan). The 401(k) Plan covers all employees who meet defined minimum age and service requirements and allows participants to defer a portion of their annual compensation on a pretax basis. The Company will make a contribution of 4% of each employee’s annual compensation up to annual safe harbor limits.
The Company has expensed $0.2 million and $0.3 million in 401(k) contributions for the six months ended June 30, 2025 and 2026, respectively.
CITY THERAPEUTICS, INC.
NOTES TO CONDENSED CONSOLIDATED FINANCIAL STATEMENTS (Continued)
(UNAUDITED)
14. Income Taxes
For the six months ended June 30, 2025 and 2026, the Company did not recognize any current or deferred income tax provision or benefit and is in a full valuation allowance. The Company did not make any federal, state, local or foreign income tax payments, net of refunds received, during the six months ended June 30, 2025 and 2026.
The Company has evaluated the positive and negative evidence bearing upon its ability to realize its deferred tax assets. Management has considered the Company’s history of cumulative net losses, estimated future taxable income, and prudent and feasible tax planning strategies and has concluded that it is more likely than not that the Company will not realize the benefits of its net deferred tax assets.
Accordingly, a full valuation allowance has been established against these net deferred tax assets as of December 31, 2025 and June 30, 2026.
15. Segments
The Company’s operations are all in the United States. The Company operates and manages its business as a single operating and reportable segment. Factors considered in determining operating and reportable segments include the organization of the business, the nature of technology and the information reviewed by the Company’s chief operating decision maker (CODM). The Company’s CODM is our CEO.
Operating segments are defined as components of an enterprise for which discrete financial information is available and is evaluated regularly by the CODM on a condensed consolidated basis, in deciding how to allocate resources and assess performance. The Company’s revenues are derived from research collaboration and license agreements, which are further described in Note 9 — Significant Agreements. Condensed consolidated net loss is used by the CODM to make key operating decisions, such as the determination of the rate at which the Company seeks to grow its research and development initiatives and the allocation of capital between research and development and general and administrative expenses. The CODM does not receive asset information other than what is presented on the condensed consolidated balance sheets.
The following table presents financial information, including significant segment expenses, which are regularly provided to the CODM and included within condensed consolidated net loss (in thousands):
| |
|
|
Six months ended
June 30,
|
|
| |
|
|
2025
|
|
|
2026
|
|
|
Collaboration revenue:
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
Ocular (Bausch + Lomb)
|
|
|
|
$ |
3,567 |
|
|
|
|
$ |
2,845 |
|
|
|
CNS (Biogen)
|
|
|
|
|
2,011 |
|
|
|
|
|
3,513 |
|
|
|
Total collaboration revenue
|
|
|
|
|
5,578 |
|
|
|
|
|
6,358 |
|
|
|
Research and development expenses:
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
CITY-FXI
|
|
|
|
|
6,217 |
|
|
|
|
|
5,565 |
|
|
|
CITY-RBP4
|
|
|
|
|
2,519 |
|
|
|
|
|
6,337 |
|
|
|
CITY-TFR2
|
|
|
|
|
516 |
|
|
|
|
|
3,543 |
|
|
|
Ocular
|
|
|
|
|
1,053 |
|
|
|
|
|
734 |
|
|
|
CNS
|
|
|
|
|
1,115 |
|
|
|
|
|
1,182 |
|
|
CITY THERAPEUTICS, INC.
NOTES TO CONDENSED CONSOLIDATED FINANCIAL STATEMENTS (Continued)
(UNAUDITED)
15. Segments (Continued)
| |
|
|
Six months ended
June 30,
|
|
| |
|
|
2025
|
|
|
2026
|
|
|
Undisclosed programs
|
|
|
|
|
— |
|
|
|
|
|
1,409 |
|
|
|
Personnel-related research and development expenses (including stock-based compensation expense of $442 and $1,316 for 2025 and 2026, respectively)
|
|
|
|
|
5,168 |
|
|
|
|
|
11,071 |
|
|
|
Other indirect research and development expenses(1)
|
|
|
|
|
7,787 |
|
|
|
|
|
16,478 |
|
|
|
Total research and development expenses
|
|
|
|
|
24,375 |
|
|
|
|
|
46,319 |
|
|
|
General and administrative expenses:
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
Personnel-related expenses (including stock-based compensation expense of
$1,055 and $3,601 for 2025 and 2026, respectively)
|
|
|
|
|
3,174 |
|
|
|
|
|
5,800 |
|
|
|
Other general and administrative expenses(2)
|
|
|
|
|
4,622 |
|
|
|
|
|
5,370 |
|
|
|
Total general and administrative expenses
|
|
|
|
|
7,796 |
|
|
|
|
|
11,170 |
|
|
|
Other segment items, net(3)
|
|
|
|
|
(2,311) |
|
|
|
|
|
2,312 |
|
|
|
Net loss and comprehensive loss
|
|
|
|
$ |
(24,282) |
|
|
|
|
$ |
(53,443) |
|
|
| |
(1)
Other indirect research and development expenses consist of platform research costs, and laboratory supplies, equipment, facility-related expenses and other general research costs that are not specifically attributable to an individual research program.
(2)
Other general and administrative expenses consist of professional fees, facilities, depreciation, information technology and other general and administrative costs that are not personnel-related.
(3)
Other segment items, net consists of interest income, interest expense and other expense, net.
16. Net Loss Per Share
Under the two-class method, basic net loss per share is calculated by dividing the net loss attributable to common stockholders by the weighted average number of shares of common stock outstanding during the period, without consideration for potentially dilutive securities. The net loss available to common stockholders was not allocated to the convertible preferred stock as the holders did not have a contractual obligation to share in losses. Diluted net loss per share is the same as basic net loss per share for the periods presented since the effects of any potentially dilutive securities would be antidilutive given the net loss of the Company.
| |
|
|
Six months ended June 30,
|
|
| |
|
|
2025
|
|
|
2026
|
|
|
Numerator:
|
|
|
|
|
Net loss attributable to common stockholders – basic and diluted
|
|
|
|
$ |
(24,282) |
|
|
|
|
$ |
(53,443) |
|
|
|
Denominator:
|
|
|
|
|
Weighted-average common shares outstanding – basic and diluted
|
|
|
|
|
4,832,880 |
|
|
|
|
|
7,096,572 |
|
|
|
Net loss per common share – basic and diluted
|
|
|
|
$ |
(5.02) |
|
|
|
|
$ |
(7.53) |
|
|
For accounting purposes, the computation of basic and diluted weighted-average common shares outstanding for the six months ended June 30, 2025 and 2026, excludes all shares of unvested restricted common stock as such shares are not considered outstanding. See Note 12 — Stock-Based Compensation, for more information.
CITY THERAPEUTICS, INC.
NOTES TO CONDENSED CONSOLIDATED FINANCIAL STATEMENTS (Continued)
(UNAUDITED)
16. Net Loss Per Share (Continued)
Correction to Previously Issued Financial Statements
The Company previously did not include shares in the basic and diluted weighted-average shares outstanding related to certain early-exercised performance-based share awards that vested during the six months ended June 30, 2026. As a result of the correction, there was an increase to the weighted-average shares outstanding for this period. The Company assessed the materiality of the change in the calculation of net loss per share, considering both quantitative and qualitative factors, and concluded that the effects of the change to the calculation and presentation of net loss per share are not material to six months ended June 30, 2026. However, the Company has revised its presentation of net loss per share previously included in the Company’s condensed consolidated financial statements for the correction of this error. All amounts have been updated to reflect the effects of the correction in the financial statements and related notes, as applicable. The following table summarizes the impact of the revision on both the weighted-average shares outstanding and net loss per share for the six months ended June 30, 2026:
| |
|
|
As reported
|
|
|
Adjustment
|
|
|
As revised
|
|
|
Impact of
stock split
|
|
|
As adjusted
for stock split
|
|
|
Weighted-average shares outstanding – basic and diluted
|
|
|
|
|
6,160,990 |
|
|
|
|
|
300,454 |
|
|
|
|
|
6,461,444 |
|
|
|
|
|
635,128 |
|
|
|
|
|
7,096,572 |
|
|
|
Net loss per common share – basic and diluted
|
|
|
|
$ |
(8.67) |
|
|
|
|
$ |
0.40 |
|
|
|
|
$ |
(8.27) |
|
|
|
|
$ |
0.74 |
|
|
|
|
$ |
(7.53) |
|
|
The following outstanding potentially dilutive securities have been excluded from the calculation of diluted net loss per share attributable to common stockholders, as their effect is antidilutive:
| |
|
|
Six months ended
June 30,
|
|
| |
|
|
2025
|
|
|
2026
|
|
|
Series Seed convertible preferred stock (as-converted basis)
|
|
|
|
|
4,991,760 |
|
|
|
|
|
4,991,760 |
|
|
|
Series A convertible preferred stock (as-converted basis)
|
|
|
|
|
15,372,145 |
|
|
|
|
|
15,372,145 |
|
|
|
Series B convertible preferred stock (as-converted basis)
|
|
|
|
|
— |
|
|
|
|
|
7,469,874 |
|
|
|
Series B-1 convertible preferred stock (as-converted basis)
|
|
|
|
|
— |
|
|
|
|
|
1,014,862 |
|
|
|
Unvested restricted stock
|
|
|
|
|
2,081,638 |
|
|
|
|
|
1,964,421 |
|
|
|
Stock options issued and outstanding
|
|
|
|
|
4,768,315 |
|
|
|
|
|
5,404,513 |
|
|
|
Total
|
|
|
|
|
27,213,858 |
|
|
|
|
|
36,217,575 |
|
|
The potentially dilutive securities related to the Biogen Note and B+L Note were excluded from the table above for the six months ended June 30, 2025 and the potentially dilutive securities related to the Biogen Note were excluded from the table above for the six months ended June 30, 2026, as the number of shares that would be issuable upon conversion was indeterminable. See Note 5 — Convertible Notes, for more information.
CITY THERAPEUTICS, INC.
NOTES TO CONDENSED CONSOLIDATED FINANCIAL STATEMENTS (Continued)
(UNAUDITED)
17. Related Party Transactions
Alexandria Real Estate Equities, Inc.
For the six months ended June 30, 2025 and 2026, the Company recognized $2.2 million and $4.4 million, respectively, in rent and facility-related fees to a subsidiary of Alexandria Real Estate Equities, Inc., a related party, in connection with leasing its headquarters at 399 Binney Street. No payments were made to Alexandria Venture Investments, LLC, an investor in the Company, holding shares of Series A convertible preferred stock for the six months ended June 30, 2025 and 2026.
Company Founder Consulting Agreement
During the six months ended June 30, 2025 and 2026, the Company recognized expenses of $0.1 million and $0.1 million, respectively, to one of the Company’s founders, who is a member of the Company’s Board, for consulting services.
18. Subsequent Events
The Company has evaluated subsequent events from the condensed balance sheet date through August 21, 2026, the date at which the condensed consolidated financial statements were available to be issued, and through September 24, 2026 and October 9, 2026, the dates the condensed consolidated financial statements were available to be reissued, and determined there are no events subject to additional disclosure, other than those disclosed below.
Option Grants
During the period from July 1, 2026 through August 21, 2026, the Company granted an aggregate of 988,458 stock options to employees and non-employee consultants under the 2023 Plan with exercise prices ranging from $7.89 to $9.10. The Company also granted an aggregate of 109,828 shares of restricted common stock to non-employee consultants under the 2023 Plan. The Company is evaluating the accounting impact of these grants.
Stock Split
On October 8, 2026, the Company effected a 1.0983-for-one stock split of its issued and outstanding shares of common stock and a proportional adjustment to the existing conversion ratios of each series of the Company’s convertible preferred stock (see Note 10). Accordingly, all share and per share amounts for all periods presented in the accompanying consolidated financial statements and notes thereto have been adjusted retroactively, where applicable, to reflect this stock split and adjustment of the preferred stock conversion ratios.
2026 Stock Option and Incentive Plan
On September 19, 2026, the Company’s Board adopted, and on October 8, 2026, the Company’s stockholders approved, the 2026 Stock Option and Incentive Plan (2026 Plan), which will become effective on the date immediately prior to the effectiveness of the registration statement for the Company’s IPO. The 2026 Plan provides for the grant of incentive stock options, nonqualified stock options, restricted stock awards, and other stock-based awards. A total of 6,516,101 shares of common stock were initially reserved for issuance under the 2026 Plan. The number of shares reserved and available for issuance under the 2026 Plan will automatically increase on January 1, 2027 and each January thereafter during the term of the 2026 Plan, by 5% of the outstanding number of shares of common stock on the immediately preceding December 31 (including, for this purpose, shares underlying any outstanding prefunded warrants) or such lesser number of shares as determined by the administrator of
CITY THERAPEUTICS, INC.
NOTES TO CONDENSED CONSOLIDATED FINANCIAL STATEMENTS (Continued)
(UNAUDITED)
18. Subsequent Events (Continued)
the 2026 Plan. Upon the effectiveness of the 2026 Plan, no further grants will be made under the 2023 Plan. The shares of common stock underlying any awards under the 2026 Plan or the 2023 Plan that are forfeited, cancelled, held back upon exercise or settlement of an award to satisfy the exercise price or tax withholding, reacquired by the Company prior to vesting, satisfied without the issuance of stock, or are otherwise terminated (other than by exercise) will be added back to the shares of common stock available for issuance under the 2026 Plan. Upon the effectiveness of the 2026 Plan, no further grants will be made under the 2023 Plan.
2026 Employee Stock Purchase Plan
On September 19, 2026, the Company’s Board adopted, and on October 8, 2026, the Company’s stockholders approved, the 2026 Employee Stock Purchase Plan (ESPP). A total of 503,322 shares of common stock were initially reserved for issuance under the ESPP. The ESPP provides that the number of shares reserved and available for issuance will automatically increase on January 1, 2027 and each January 1 thereafter through January 1, 2036, by the least of (i) 1% of the outstanding number of shares of common stock on the immediately preceding December 31 (including, for this purpose, shares underlying any outstanding prefunded warrants), (ii) 1,509,966 shares of common stock, or (iii) such number of shares of common stock as determined by the administrator of the ESPP.
Shares
Common Stock
Preliminary Prospectus
| |
Goldman Sachs & Co. LLC
|
|
|
Jefferies
|
|
|
Stifel
|
|
| |
Oppenheimer & Co.
|
|
Until , 2026 (25 days after the date of this prospectus), all dealers that effect transactions in these securities, whether or not participating in this offering, may be required to deliver a prospectus. This delivery requirement is in addition to the dealers’ obligation to deliver a prospectus when acting as underwriters and with respect to their unsold allotments or subscriptions.
, 2026
PART II
INFORMATION NOT REQUIRED IN PROSPECTUS
Unless otherwise indicated, all references to “City,” “City Therapeutics,” the “company,” “we,” “our,” “us” or similar terms refer to City Therapeutics, Inc. and its wholly owned subsidiary, or either or both of them as the context may require.
Item 13. Other Expenses of Issuance and Distribution.
The following table sets forth all expenses to be paid by us, other than underwriting discounts and commissions, in connection with this offering. All amounts shown are estimates except for the Securities and Exchange Commission (“SEC”) registration fee, the Financial Industry Regulatory Authority, Inc. (“FINRA”) filing fee and The Nasdaq Stock Market (“Nasdaq”) listing fee.
| |
SEC registration fee
|
|
|
|
$ |
23,591 |
|
|
| |
FINRA filing fee
|
|
|
|
|
32,365 |
|
|
| |
Nasdaq listing fee
|
|
|
|
|
325,000 |
|
|
| |
Printing and mailing expenses
|
|
|
|
|
250,000 |
|
|
| |
Legal fees and expenses
|
|
|
|
|
2,750,000 |
|
|
| |
Accounting fees and expenses
|
|
|
|
|
950,000 |
|
|
| |
Custodian transfer agent and registrar fees and expenses
|
|
|
|
|
10,000 |
|
|
| |
Miscellaneous expenses
|
|
|
|
|
159,044 |
|
|
| |
Total
|
|
|
|
$ |
4,500,000 |
|
|
*
To be provided by amendment.
Item 14. Indemnification of Directors and Officers.
Section 145 of the Delaware General Corporation Law (the “DGCL”) authorizes a corporation to indemnify its directors and officers against liabilities arising out of actions, suits and proceedings to which they are made or threatened to be made a party by reason of the fact that they have served or are currently serving as a director or officer to a corporation. The indemnity may cover expenses (including attorneys’ fees) judgments, fines, and amounts paid in settlement actually and reasonably incurred by the director or officer in connection with any such action, suit or proceeding. Section 145 permits corporations to pay expenses (including attorneys’ fees) incurred by directors and officers in advance of the final disposition of such action, suit or proceeding. In addition, Section 145 provides that a corporation has the power to purchase and maintain insurance on behalf of its directors and officers against any liability asserted against them and incurred by them in their capacity as a director or officer, or arising out of their status as such, whether or not the corporation would have the power to indemnify the director or officer against such liability under Section 145.
We will adopt provisions in our amended and restated certificate of incorporation, which will become effective immediately prior to the completion of this offering, and the amended and restated bylaws, which will become effective upon the effectiveness of the registration statement of which this prospectus forms a part, that limit or eliminate the personal liability of our directors and officers to the fullest extent permitted by the DGCL, as it now exists or may in the future be amended. Consequently, our directors and officers will not be personally liable to us or our stockholders for monetary damages or breach of fiduciary duty as directors or officers, except for liability for:
•
any breach of their duty of loyalty to us or our stockholders;
•
any act or omission not in good faith or that involves intentional misconduct or a knowing violation of law;
•
for our directors, any unlawful payments related to dividends or unlawful stock purchases, redemptions or other distributions;
•
any transaction from which they derived an improper personal benefit; or
•
for our officers, any derivative action by or in the right of the corporation.
These limitations of liability do not alter director and officer liability under the federal securities laws and do not affect the availability of equitable remedies such as an injunction or rescission.
In addition, our amended and restated bylaws will provide that:
•
we will indemnify our directors, officers and, in the discretion of our board of directors, certain employees to the fullest extent permitted by the DGCL, as it now exists or may in the future be amended; and
•
we will advance reasonable expenses, including attorneys’ fees, to our directors and, in the discretion of our board of directors, to our officers and certain employees, in connection with legal proceedings relating to their service for or on behalf of us, subject to limited exceptions.
We have entered into or intend to enter into indemnification agreements with each of our directors and executive officers. These agreements provide that we will indemnify each of our directors, our executive officers and, at times, their affiliates to the fullest extent permitted by Delaware law. We will advance expenses, including attorneys’ fees (but excluding judgments, fines and settlement amounts), to each indemnified director, executive officer or affiliate in connection with any proceeding in which indemnification is available and we will indemnify our directors and officers for any action or proceeding arising out of that person’s services as a director or officer brought on behalf of us or in furtherance of our rights. Additionally, certain of our directors or officers may have certain rights to indemnification, advancement of expenses or insurance provided by their affiliates or other third parties, which indemnification relates to and might apply to the same proceedings arising out of such director’s or officer’s services as a director referenced herein. Nonetheless, we have agreed in the indemnification agreements that our obligations to those same directors or officers are primary and any obligation of such affiliates or other third parties to advance expenses or to provide indemnification for the expenses or liabilities incurred by those directors are secondary.
We also maintain general liability insurance which covers certain liabilities of our directors and officers arising out of claims based on acts or omissions in their capacities as directors or officers, including liabilities under the Securities Act of 1933, as amended (the “Securities Act”).
The underwriting agreement filed as Exhibit 1.1 to this registration statement provides for indemnification of us and our directors and officers by the underwriters against certain liabilities under the Securities Act and the Securities Exchange Act of 1934, as amended.
Item 15. Recent Sales of Unregistered Securities.
Set forth below is information regarding securities we have issued within the past three years that were not registered under the Securities Act.
(a) Convertible Promissory Notes
From July 2024 through May 2025, we issued convertible promissory notes with an aggregate principal amount of $40.0 million.
(b) Issuances of Capital Stock
Preferred Stock Issuances
From September 2023 through March 2024, we issued and sold to certain accredited investors an aggregate of 4,544,990 shares of our Series Seed convertible preferred stock at a purchase price of $1.00 per share, for aggregate gross proceeds of approximately $4.5 million.
From April 2024 through December 2024, we issued and sold to certain accredited investors an aggregate of 13,435,354 shares of our Series A convertible preferred stock at a purchase price of
$10.0295 per share, for aggregate gross proceeds of approximately $134.7 million. During this period, we also issued an aggregate of 560,964 shares of our Series A convertible preferred stock upon the conversion of outstanding convertible promissory notes and related accrued interest at a conversion price of $9.02655 per share.
From May 2026 through June 2026, we issued and sold to certain accredited investors an aggregate of 6,801,318 shares of our Series B convertible preferred stock at a purchase price of $14.6295 per share, for aggregate gross proceeds of approximately $99.5 million. During this period, we also issued an aggregate of 924,030 shares of our Series B convertible preferred stock upon the conversion of outstanding convertible promissory notes and related accrued interest at a conversion price of $12.4351 per share.
Common Stock Issuances
Since August 2023, we have issued and sold by exchange to our founders an aggregate of 5,842,014 shares of restricted common stock pursuant to stock purchase agreements for an insignificant amount of cash consideration.
From October 2023 through April 2024, we issued an aggregate of 655,225 shares of our common stock to the Ohio State Innovation Foundation in connection with the exclusive license agreement.
In July 2026, we issued and sold by exchange to certain of our consultants an aggregate of 109,828 shares of restricted common stock pursuant to stock purchase agreements for an insignificant amount of cash consideration.
The offers, sales and issuances of the securities described above were deemed to be exempt under Section 4(a)(2) of the Securities Act or Rule 506 of Regulation D under the Securities Act as a transaction by an issuer not involving a public offering. The recipients of securities in each of these transactions acquired the securities for investment only and not with a view to or for sale in connection with any distribution thereof and appropriate legends were affixed to the securities issued in these transactions. Each of the recipients of securities in these transactions was an accredited investor within the meaning of Rule 501 of Regulation D under the Securities Act and had adequate access, through employment, business or other relationships, to information about us. No underwriters were involved in these transactions.
(c) Grants and Exercises of Stock Options
Since February 2024, we have granted certain of our employees, consultants, and directors options to purchase an aggregate of 9,637,337 shares of our common stock under our 2023 Stock Plan, as amended from time to time (the “2023 Plan”), at exercise prices ranging from $0.08 to $9.10 per share.
Since January 2025, options to purchase an aggregate of 3,219,530 shares of our common stock under our 2023 Plan have been exercised at a weighted average exercise price of $0.98 per share.
The offers, sales and issuances of the securities described above were deemed to be exempt from registration under Rule 701 promulgated under the Securities Act as transactions under compensatory benefit plans and contracts relating to compensation, or under Section 4(a)(2) of the Securities Act as a transaction by an issuer not involving a public offering. The recipients of such securities were our directors, employees or bona fide consultants and received the securities under our equity incentive plans. Appropriate legends were affixed to the securities issued in these transactions. Each of the recipients of securities in these transactions had adequate access, through employment, business or other relationships, to information about us.
Item 16. Exhibits and Financial Statement Schedules.
(a) Exhibits.
| |
Exhibit
Number
|
|
|
Description
|
|
| |
1.1
|
|
|
|
|
| |
3.1
|
|
|
|
|
| |
3.2
|
|
|
|
|
| |
3.3*
|
|
|
|
|
| |
3.4
|
|
|
|
|
| |
4.1*
|
|
|
|
|
| |
4.2+*
|
|
|
|
|
| |
5.1
|
|
|
|
|
| |
10.1#*
|
|
|
|
|
| |
10.2#
|
|
|
|
|
| |
10.3#
|
|
|
|
|
| |
10.4#*
|
|
|
|
|
| |
10.5#*
|
|
|
|
|
| |
10.6#
|
|
|
|
|
| |
10.7#
|
|
|
|
|
| |
10.8#
|
|
|
|
|
| |
10.9#
|
|
|
|
|
| |
10.10†+*
|
|
|
|
|
| |
10.11†+*
|
|
|
|
|
| |
10.12†+*
|
|
|
Exclusive License Agreement, by and between the Ohio State Innovation Foundation and the Registrant, dated as of October 30, 2023, as amended by Amendment No. 1, dated as of February 8, 2024, Amendment No. 2, dated as of April 3, 2025, Amendment No. 3, dated as of May 13, 2025, Amendment No. 4, dated as of July 4, 2025, and Amendment No. 5, dated as of August 26, 2025.
|
|
| |
10.13#+*
|
|
|
|
|
| |
10.14#*
|
|
|
|
|
| |
10.15#+*
|
|
|
|
|
| |
10.16#+*
|
|
|
|
|
| |
Exhibit
Number
|
|
|
Description
|
|
| |
10.17#+*
|
|
|
|
|
| |
10.18+*
|
|
|
|
|
| |
21.1*
|
|
|
|
|
| |
23.1
|
|
|
|
|
| |
23.2
|
|
|
|
|
| |
24.1*
|
|
|
|
|
| |
107
|
|
|
|
|
*
Previously filed.
#
Indicates a management contract or any compensatory plan, contract or arrangement.
†
Certain portions of this document that constitute confidential information have been redacted pursuant to Item 601(b)(10) of Regulation S-K.
+
Certain exhibits and schedules to these agreements have been omitted pursuant to Item 601(a)(5) and (6) of Regulation S-K. The registrant will furnish copies of any of the exhibits and schedules to the Securities and Exchange Commission upon request.
(b) Financial Statement Schedules.
All financial statement schedules are omitted because the information required to be set forth therein is not applicable or is shown in the audited consolidated financial statements or the notes thereto.
Item 17. Undertakings.
Insofar as indemnification for liabilities arising under the Securities Act may be permitted to directors, officers and controlling persons of the registrant pursuant to the foregoing provisions, or otherwise, the registrant has been advised that in the opinion of the SEC such indemnification is against public policy as expressed in the Securities Act and is, therefore, unenforceable. In the event that a claim for indemnification against such liabilities (other than the payment by the registrant of expenses incurred or paid by a director, officer or controlling person of the registrant in the successful defense of any action, suit or proceeding) is asserted by such director, officer or controlling person in connection with the securities being registered, the registrant will, unless in the opinion of its counsel the matter has been settled by controlling precedent, submit to a court of appropriate jurisdiction the question whether such indemnification by it is against public policy as expressed in the Securities Act and will be governed by the final adjudication of such issue.
The registrant hereby undertakes that:
(a)
The Registrant will provide to the underwriter at the closing as specified in the underwriting agreement, certificates in such denominations and registered in such names as required by the underwriter to permit prompt delivery to each purchaser.
(b)
For purposes of determining any liability under the Securities Act, as amended, the information omitted from a form of prospectus filed as part of this registration statement in reliance upon Rule 430A and contained in the form of prospectus filed by the Registrant pursuant to Rule 424(b)(1) or (4) or 497(h) under the Securities Act, as amended, shall be deemed to be part of this registration statement as of the time it was declared effective.
(c)
For the purpose of determining any liability under the Securities Act, as amended, each post-effective amendment that contains a form of prospectus shall be deemed to be a new registration statement relating to the securities offered therein, and the offering of such securities at that time shall be deemed to be the initial bona fide offering thereof.
SIGNATURES
Pursuant to the requirements of the Securities Act of 1933, as amended, the registrant has duly caused this Amendment No. 1 to the Registration Statement on Form S-1 to be signed on its behalf by the undersigned, thereunto duly authorized, in the City of Cambridge, Massachusetts, on the 9th of October, 2026.
CITY THERAPEUTICS, INC.
By
/s/ Andrew Orth, M.B.A.
Name:
Andrew Orth, M.B.A.
Title:
President and Chief Executive Officer
| |
Signature
|
|
|
Title
|
|
|
Date
|
|
| |
/s/ Andrew Orth, M.B.A.
Andrew Orth, M.B.A.
|
|
|
President, Chief Executive Officer and Director (Principal Executive Officer)
|
|
|
October 9, 2026
|
|
| |
/s/ Keith Regnante, M.B.A.
Keith Regnante, M.B.A.
|
|
|
Chief Financial Officer (Principal Financial and Accounting Officer)
|
|
|
October 9, 2026
|
|
| |
*
John Maraganore, Ph.D.
|
|
|
Executive Chair and Director
|
|
|
October 9, 2026
|
|
| |
*
Barry Greene
|
|
|
Director
|
|
|
October 9, 2026
|
|
| |
*
Robert Nelsen, M.B.A.
|
|
|
Director
|
|
|
October 9, 2026
|
|
| |
*
Saraswathy V. Nochur, Ph.D.
|
|
|
Director
|
|
|
October 9, 2026
|
|
| |
*
Ron Philip
|
|
|
Director
|
|
|
October 9, 2026
|
|
*
By: /s/ Andrew Orth, M.B.A.
Andrew Orth, M.B.A.
Attorney-In-Fact
Exhibit 1.1
City Therapeutics, Inc.
Common Stock,
par value $0.0001 per share
Underwriting Agreement
[·], 2026
Goldman Sachs & Co. LLC
Jefferies LLC
As representatives (the “Representatives”)
of the several Underwriters
named in Schedule I hereto,
c/o Goldman
Sachs & Co. LLC
200 West Street
New York, New York 10282
c/o Jefferies
LLC
520 Madison Avenue
New York, New York 10022
Ladies and Gentlemen:
City Therapeutics, Inc.,
a Delaware corporation (the “Company”), proposes, subject to the terms and conditions stated in this agreement (this “Agreement”),
to issue and sell to the Underwriters named in Schedule I hereto (the “Underwriters”) an aggregate of [·] shares (the
“Firm Shares”) and, at the election of the Underwriters, up to [·] additional shares (the “Optional Shares”)
of common stock, par value $0.0001 per share (“Stock”) of the Company. The Firm Shares and the Optional Shares that the Underwriters
elect to purchase pursuant to Section 2 hereof are herein collectively called the “Shares”.
1. The
Company represents and warrants to, and agrees with, each of the Underwriters that:
(a) A
registration statement on Form S-1 (File No. 333-299117) (the “Initial Registration Statement”) in respect of the
Shares has been filed with the Securities and Exchange Commission (the “Commission”); the Initial Registration Statement and
any post-effective amendment thereto, each in the form heretofore delivered to you, have been declared effective by the Commission in
such form; other than a registration statement, if any, increasing the size of the offering (a “Rule 462(b) Registration
Statement”), filed pursuant to Rule 462(b) under the Securities Act of 1933, as amended (the “Act”), which
became effective upon filing, no other document with respect to the Initial Registration Statement has been filed with the Commission;
and no stop order suspending the effectiveness of the Initial Registration Statement, any post-effective amendment thereto or the Rule 462(b) Registration
Statement, if any, has been issued and no proceeding for that purpose or pursuant to Section 8A of the Act has been initiated or,
to the Company’s knowledge, threatened by the Commission (any preliminary prospectus included in the Initial Registration Statement
or filed with the Commission pursuant to Rule 424(a) of the rules and regulations of the Commission under the Act is hereinafter
called a “Preliminary Prospectus”; the various parts of the Initial Registration Statement and the Rule 462(b) Registration
Statement, if any, including all exhibits thereto and including the information contained in the form of final prospectus filed with the
Commission pursuant to Rule 424(b) under the Act in accordance with Section 5(a) hereof and deemed by virtue of Rule 430A
under the Act to be part of the Initial Registration Statement at the time it was declared effective, each as amended at the time such
part of the Initial Registration Statement became effective or such part of the Rule 462(b) Registration Statement, if any,
became or hereafter becomes effective, are hereinafter collectively called the “Registration Statement”; the Preliminary Prospectus
relating to the Shares that was included in the Registration Statement immediately prior to the Applicable Time (as defined in Section 1(c) hereof)
is hereinafter called the “Pricing Prospectus”; such final prospectus, in the form first filed pursuant to Rule 424(b) under
the Act, is hereinafter called the “Prospectus”; any oral or written communication with potential investors undertaken in
reliance on Section 5(d) of the Act or Rule 163B under the Act is hereinafter called a “Testing-the-Waters Communication”;
and any Testing-the-Waters Communication that is a written communication within the meaning of Rule 405 under the Act is hereinafter
called a “Written Testing-the-Waters Communication”; and any “issuer free writing prospectus” as defined in Rule 433
under the Act relating to the Shares is hereinafter called an “Issuer Free Writing Prospectus”); and any “bona fide
electronic road show” as defined in Rule 433(h)(5) under the Act that has been made available without restriction to any
person is hereinafter called a “broadly available road show”;
(b) (A) No
order preventing or suspending the use of any Preliminary Prospectus or any Issuer Free Writing Prospectus has been issued by the Commission,
(B) no proceeding for that purpose or pursuant to Section 8A of the Securities Act against the Company or related to the offering
of the Shares has been initiated or to the Company’s knowledge, threatened by the Commission, and (C) each Preliminary Prospectus,
at the time of filing thereof, conformed in all material respects to the requirements of the Act and the rules and regulations of
the Commission thereunder, and did not contain an untrue statement of a material fact or omit to state a material fact required to be
stated therein or necessary to make the statements therein, in the light of the circumstances under which they were made, not misleading;
provided, however, that this representation and warranty shall not apply to any statements or omissions made in reliance upon and in conformity
with the Underwriter Information (as defined in Section 9(b) of this Agreement);
(c) For
the purposes of this Agreement, the “Applicable Time” is [·] p.m., New York time, on the date of this Agreement. The
Pricing Prospectus, as supplemented by the information listed on Schedule II(c) hereto, taken together (collectively, the “Pricing
Disclosure Package”), as of the Applicable Time, did not, and as of each Time of Delivery (as defined in Section 4(a) of
this Agreement), will not, include any untrue statement of a material fact or omit to state any material fact necessary in order to make
the statements therein, in the light of the circumstances under which they were made, not misleading; and each Issuer Free Writing Prospectus,
each broadly available road show and each Written Testing-the-Waters Communication does not conflict with the information contained in
the Registration Statement, the Pricing Prospectus or the Prospectus, and each Issuer Free Writing Prospectus, each broadly available
road show and each Written Testing-the-Waters Communication, as supplemented by and taken together with the Pricing Disclosure Package,
as of the Applicable Time, did not, and as of each Time of Delivery, will not, include any untrue statement of a material fact or omit
to state any material fact necessary in order to make the statements therein, in the light of the circumstances under which they were
made, not misleading; provided, however, that this representation and warranty shall not apply to any statements or omissions made in
reliance upon and in conformity with the Underwriter Information;
(d) No
documents were filed by the Company with the Commission since the Commission’s close of business on the business day immediately
prior to the date of this Agreement and prior to the execution of this Agreement, except as set forth on Schedule II(b) hereto;
(e) The
Registration Statement conforms, and the Prospectus and any further amendments or supplements to the Registration Statement and the Prospectus
will conform, in all material respects to the requirements of the Act and the rules and regulations of the Commission thereunder
and do not and will not, as of the applicable effective date as to each part of the Registration Statement, as of the applicable filing
date as to the Prospectus and any amendment or supplement thereto, and as of each Time of Delivery, contain an untrue statement of a material
fact or omit to state a material fact required to be stated therein or necessary to make the statements therein, in the case of the Prospectus,
in light of the circumstances under which they were made, not misleading; provided, however, that this representation and warranty shall
not apply to any statements or omissions made in reliance upon and in conformity with the Underwriter Information;
(f) Neither
the Company nor any of its subsidiaries has, since the date of the latest audited financial statements included in the Pricing Prospectus,
(i) sustained any material loss or material interference with its business from fire, explosion, flood or other calamity, whether
or not covered by insurance, or from any labor dispute or court or governmental action, order or decree or (ii) entered into any
transaction or agreement (whether or not in the ordinary course of business) that is material to the Company and its subsidiaries taken
as a whole or incurred any liability or obligation, direct or contingent, that is material to the Company and its subsidiaries taken as
a whole, in each case otherwise than as set forth or contemplated in the Pricing Prospectus; and, since the respective dates as of which
information is given in the Registration Statement and the Pricing Prospectus, there has not been (A) any change in the capital stock
(other than as a result of (1) the exercise, if any, of stock options or warrants or settlement of restricted stock units (including
any “net” or “cashless” exercises or settlements), if any, or the award, if any, of stock options, restricted
stock units, restricted stock or other awards pursuant to the Company’s equity plans that are described in the Pricing Prospectus
and the Prospectus, (2) the repurchase of shares of capital stock upon termination of the holder’s employment or service with
the Company pursuant to agreements providing for an option to repurchase or a right of first refusal on behalf of the Company, (3) the
issuance, if any, of stock upon exercise, conversion, exchange or reclassification of Company securities as described in the Pricing Prospectus
and the Prospectus, or (4) issuances otherwise set forth or contemplated in the Pricing Prospectus), (B) any increase in long
term debt of the Company or any of its subsidiaries or (C) any Material Adverse Effect (as defined below); as used in this Agreement,
“Material Adverse Effect” shall mean any material adverse change or effect, or any development involving a prospective material
adverse change or effect, in or affecting (1) the business, properties, general affairs, management, financial position, stockholders’
deficit or results of operations of the Company and its subsidiaries, taken as a whole, except as set forth or contemplated in the Pricing
Prospectus, or (2) the ability of the Company to perform its obligations under this Agreement, including the issuance and sale of
the Shares, or to consummate the transactions contemplated in the Pricing Prospectus and the Prospectus;
(g) The
Company and its subsidiaries have good and marketable title in fee simple to all real property and good and marketable title to all personal
property owned by them, in each case free and clear of all liens, encumbrances and defects except such as do not materially affect the
value of such property and do not materially interfere with the use made and proposed to be made of such property by the Company and its
subsidiaries; and any real property and buildings held under lease by the Company and its subsidiaries are held by them under valid, subsisting
and enforceable leases with such exceptions as are not material and do not materially interfere with the use made and proposed to be made
of such property and buildings by the Company and its subsidiaries;
(h) Each
of the Company and each of its subsidiaries has been (i) duly organized and is validly existing and in good standing under the laws
of its jurisdiction of organization, with power and authority (corporate and other) to own its properties and conduct its business as
described in the Pricing Prospectus, and (ii) duly qualified as a foreign corporation or other business entity for the transaction
of business and is in good standing (where such concept exists) under the laws of each other jurisdiction in which it owns or leases properties
or conducts any business so as to require such qualification, except, in the case of this clause (ii), where the failure to be so qualified
or in good standing would not, individually or in the aggregate, reasonably be expected to have a Material Adverse Effect, and each subsidiary
of the Company has been listed in Exhibit 21.1 to the Registration Statement;
(i) The
Company has an authorized capitalization as set forth in the Pricing Prospectus and all of the issued shares of capital stock of the Company
have been duly and validly authorized and issued, are fully paid and non-assessable and conform, in all material respects, to the description
of the Stock contained in the Pricing Disclosure Package and the Prospectus; and all of the issued shares of capital stock of each subsidiary
of the Company have been duly and validly authorized and issued, are fully paid and non-assessable and (except, in the case of any foreign
subsidiary, for directors’ qualifying shares) are owned directly or indirectly by the Company, free and clear of all liens, encumbrances,
equities or claims; and none of the outstanding capital stock or equity interest in any subsidiary was issued in violation of preemptive
or similar rights of any security holder of such subsidiary;
(j) The
unissued Shares to be issued and sold by the Company to the Underwriters hereunder have been duly and validly authorized and, when issued
and delivered against payment therefor as provided herein, will be duly and validly issued and fully paid and non-assessable and will
conform, in all material respects, to the description of the Stock contained in the Pricing Disclosure Package and the Prospectus; and
the issuance of the Shares is not subject to any preemptive or similar rights, except as have been duly and validly waived or complied
with;
(k) The
issue and sale of the Shares and the compliance by the Company with this Agreement and the consummation of the transactions contemplated
in this Agreement and the Pricing Prospectus will not conflict with or result in a breach or violation of any of the terms or provisions
of, or constitute a default under, (A) any indenture, mortgage, deed of trust, loan agreement, lease or other agreement or instrument
to which the Company or any of its subsidiaries is a party or by which the Company or any of its subsidiaries is bound or to which any
of the property or assets of the Company or any of its subsidiaries is subject, (B) the certificate of incorporation or by-laws (or
other applicable organizational document) of the Company or any of its subsidiaries, or (C) any statute or any judgment, order, rule or
regulation of any court or governmental agency or body having jurisdiction over the Company or any of its subsidiaries or any of their
properties, except, in the cases of clauses (A) and (C) for such defaults, breaches, or violations that would not, individually
or in the aggregate, have a Material Adverse Effect; and no consent, approval, authorization, order, registration or qualification of
or with any such court or governmental agency or body is required for the issue and sale of the Shares or the consummation by the Company
of the transactions contemplated by this Agreement, except such as have been obtained under the Act, the approval by the Financial Industry
Regulatory Authority (“FINRA”) of the underwriting terms and arrangements, the approval for listing the Shares on The Nasdaq
Stock Market (the “Exchange”) and such consents, approvals, authorizations, orders, registrations or qualifications as may
be required under state securities or Blue Sky laws in connection with the purchase and distribution of the Shares by the Underwriters;
(l) Neither
the Company nor any of its subsidiaries is (i) in violation of its certificate of incorporation or by-laws (or other applicable organizational
document), (ii) in violation of any statute or any judgment, order, rule or regulation of any court or governmental agency or
body having jurisdiction over the Company or any of its subsidiaries or any of their properties, or (iii) in default in the performance
or observance of any obligation, agreement, covenant or condition contained in any indenture, mortgage, deed of trust, loan agreement,
lease or other agreement or instrument to which it is a party or by which it or any of its properties may be bound, except, in the case
of the foregoing clauses (ii) and (iii), for such defaults as would not, individually or in the aggregate, reasonably be expected
to have a Material Adverse Effect;
(m) The
statements set forth in the Pricing Prospectus and the Prospectus under the caption “Description of Capital Stock” and “Shares
Eligible for Future Sale,” insofar as they purport to constitute a summary of the terms of the Stock and under the captions “Material
U.S. Federal Income Tax Consequences to Non-U.S. Holders” and “Underwriting”, insofar as they purport to describe the
provisions of the laws and documents referred to therein, are accurate, complete and fair in all material respects;
(n) There
are no legal, governmental or regulatory investigations, actions, demands, claims, suits, arbitrations, inquiries or proceedings (“Actions”)
pending to which the Company or any of its subsidiaries or any of their properties or assets or, to the Company’s knowledge, any
officer or director of the Company is a party or of which any property or assets of the Company or any of its subsidiaries or any of their
properties or assets or, to the Company’s knowledge, any officer or director of the Company is the subject which, if determined
adversely to the Company or any of its subsidiaries (or such officer or director), would individually or in the aggregate have a Material
Adverse Effect; and, to the Company’s knowledge, no such proceedings are threatened or contemplated by governmental authorities
or others; there are no current or pending Actions that are required under the Act to be described in the Registration Statement or the
Pricing Prospectus that are not so described therein; and there are no statutes, regulations or contracts or other documents that are
required under the Act to be filed as exhibits to the Registration Statement or described in the Registration Statement, the Pricing Prospectus
that are not so filed as exhibits to the Registration Statement or described in the Registration Statement and the Pricing Prospectus;
(o) The
Company is not and, after giving effect to the offering and sale of the Shares and the application of the proceeds thereof as described
in the Pricing Prospectus and the Prospectus, will not be an “investment company”, as such term is defined in the Investment
Company Act of 1940, as amended (the “Investment Company Act”);
(p) At
the time of filing the Initial Registration Statement and any post-effective amendment thereto, at the earliest time thereafter that the
Company or any offering participant made a bona fide offer (within the meaning of Rule 164(h)(2) under the Act) of the Shares,
and at the date hereof, the Company was not and is not an “ineligible issuer,” as defined under Rule 405 under the Act;
(q) PricewaterhouseCoopers
LLP, who have certified certain financial statements of the Company and its subsidiaries, are independent public accountants as required
by the Act and the rules and regulations of the Commission thereunder;
(r) The
Company maintains a system of internal control over financial reporting (as such term is defined in Rule 13a-15(f) under the
Securities Exchange Act of 1934, as amended (the “Exchange Act”)), that (i) complies with the requirements of the Exchange
Act applicable to the Company, (ii) has been designed by the Company’s principal executive officer and principal financial
officer, or under their supervision, to provide reasonable assurance regarding the reliability of financial reporting and the preparation
of financial statements for external purposes in accordance with U.S. generally accepted accounting principles (“GAAP”) and
(iii) is sufficient to provide reasonable assurance that (A) transactions are executed in accordance with management’s
general or specific authorization, (B) transactions are recorded as necessary to permit preparation of financial statements in conformity
with GAAP and to maintain accountability for assets, (C) access to assets is permitted only in accordance with management’s
general or specific authorization and (D) the recorded accountability for assets is compared with the existing assets at reasonable
intervals and appropriate action is taken with respect to any differences (it being understood that this subsection shall not require
the Company to comply with Section 404 of the Sarbanes-Oxley Act of 2002, as amended, and the rules and regulations promulgated
in connection therewith (the “Sarbanes-Oxley Act”), as of an earlier date than it would otherwise be required to so comply
under applicable law); and the Company’s internal control over financial reporting is effective and the Company is not aware of
any material weaknesses in its internal control over financial reporting. There are no material weaknesses in the Company’s internal
controls;
(s) Since
the date of the latest audited financial statements included in the Pricing Prospectus, there has been no change in the Company’s
internal control over financial reporting that has materially and adversely affected, or is reasonably likely to materially and adversely
affect, the Company’s internal control over financial reporting;
(t) The
Company maintains disclosure controls and procedures (as such term is defined in Rule 13a-15(e) under the Exchange Act) that
comply with the requirements of the Exchange Act applicable to the Company; such disclosure controls and procedures have been designed
to ensure that material information relating to the Company and its subsidiaries is made known to the Company’s principal executive
officer and principal financial officer by others within those entities; and such disclosure controls and procedures are effective;
(u) This
Agreement has been duly authorized, executed and delivered by the Company;
(v) The
Company has full right, power and authority to execute and deliver this Agreement and to perform its obligations hereunder; and all action
required to be taken for the due and proper authorization, execution and delivery by the Company of this Agreement and the consummation
by the Company of the transactions contemplated hereby has been duly and validly taken;
(w) (A) None
of the Company or any of its subsidiaries, or any director, officer, or employee thereof, nor, to the knowledge of the Company, any agent,
affiliate or other person associated with or acting on behalf of the Company or any of its subsidiaries or affiliates, has (i) made,
offered, promised or authorized any unlawful contribution, gift, entertainment or other unlawful expense (or taken any act in furtherance
thereof), (ii) taken or will take any action in furtherance of an offer, payment, promise to pay, or authorization or approval of
the payment, giving or receipt of money, property, gifts, entertainment, other expense or anything else of value, directly or indirectly,
to any government official (including any officer or employee of a government or government-owned or controlled entity or of a public
international organization, or any person acting in an official capacity for or on behalf of any of the foregoing, or any political party
or party official or candidate for political office) in order to influence official action, or to any person in violation of any applicable
anti-corruption laws; or (iii) violated or is in violation of any provision of the U.S. Foreign Corrupt Practices Act of 1977, as
amended, the Bribery Act 2010 of the United Kingdom or any other applicable anti-bribery or anti-corruption laws; (B) the Company
and its subsidiaries and, to the Company’s knowledge, its affiliates have conducted their businesses in compliance with applicable
anti-bribery and anti-corruption laws and have instituted and maintained, and will continue to maintain, policies and procedures reasonably
designed to promote and achieve compliance with such laws and with the representations and warranties contained herein; and (C) neither
the Company nor any of its subsidiaries will use, directly or indirectly, the proceeds of the offering of the Shares hereunder in furtherance
of any unlawful expense or any direct or indirect unlawful payment or in violation of any applicable anti-bribery or anti-corruption laws;
(x) The
operations of the Company and its subsidiaries are and have been conducted at all times in compliance with the requirements of applicable
financial recordkeeping and reporting requirements and anti-money laundering laws, including, but not limited to, the Bank Secrecy Act
of 1970, as amended by the USA PATRIOT ACT of 2001, and the rules and regulations promulgated thereunder, and any similar rules,
regulations or guidelines issued, administered or enforced by any governmental agency, of the various jurisdictions in which the Company
and its subsidiaries conduct business (collectively, the “Money Laundering Laws”) and no action, suit or proceeding by or
before any court or governmental agency, authority or body or any arbitrator involving the Company or any of its subsidiaries with respect
to the Money Laundering Laws is pending or, to the knowledge of the Company, threatened;
(y) (A) None
of the Company or any of its subsidiaries, or any director, officer or employee thereof, nor, to the knowledge of the Company, any agent,
affiliate, or other person associated with or acting on behalf of the Company or any of its subsidiaries is a person or entity that is,
or is 50% or more owned or, where relevant under applicable Sanctions (as defined below), controlled by one or more persons or entities
that are, currently the subject or the target of any sanctions administered or enforced by the U.S. government, including, without limitation,
the Office of Foreign Assets Control of the U.S. Department of the Treasury (“OFAC”), or the U.S. Department of State (and
including, without limitation, the designation as a “specially designated national” or “blocked person”), the
European Union, His Majesty’s Treasury, the United Nations Security Council, or other relevant sanctions authority (collectively,
“Sanctions”), or located, organized, or resident in a country or territory that is the subject or target of Sanctions (currently,
Crimea, Cuba, Iran, North Korea, Syria (prior to July 1, 2025), the so-called Donetsk People’s Republic and so-called
Luhansk People’s Republic, and the non-government controlled areas of Zaporizhzhia and Kherson regions of Ukraine (each a “Sanctioned
Jurisdiction”)); (B) the Company will not directly or indirectly use the proceeds of the offering of the Shares hereunder,
or lend, contribute or otherwise make available such proceeds to any subsidiary, joint venture partner or other person or entity (i) to
fund or facilitate any activities of or business with any Sanctioned Jurisdiction or any person or entity, that, at the time of such funding
or facilitation, is the subject or the target of Sanctions or (ii) in any other manner that will result in a violation by any person
or entity (including any person or entity participating in the transaction, whether as underwriter, advisor, investor or otherwise) of
Sanctions; and (C) the Company and its subsidiaries have not, since April 24, 2019, knowingly engaged in, are not now knowingly
engaged in and will not engage in any dealings or transactions with any Sanctioned Jurisdiction or involving any person or entity, that,
at the time of such dealing or transaction, is or was the subject or the target of Sanctions; the Company and its subsidiaries have instituted,
and maintain, policies and procedures designed to promote and achieve continued compliance with Sanctions;
(z) The
financial statements included in the Registration Statement, the Pricing Prospectus and the Prospectus, together with the related schedules
and notes, present fairly, in all material respects, the financial position of the Company and its subsidiaries at the dates indicated
and the statement of operations and comprehensive loss, convertible preferred stock and stockholders’ deficit and cash flows of
the Company and its subsidiaries for the periods specified; said financial statements have been prepared in conformity with GAAP applied
on a consistent basis throughout the periods involved, except in the case of any unaudited financial statements, which are subject to
normal year-end adjustments and do not contain certain footnotes as permitted by the applicable rules of the Commission. The supporting
schedules, if any, present fairly, in all material respects, in accordance with GAAP the information required to be stated therein. The
selected financial data and the summary financial information included in the Registration Statement, the Pricing Prospectus and the Prospectus
present fairly, in all material respects, the information shown therein and have been compiled on a basis consistent with that of the
audited financial statements included therein. Except as included therein, no historical or pro forma financial statements or supporting
schedules are required to be included in the Registration Statement, the Pricing Prospectus or the Prospectus under the Act or the rules and
regulations promulgated thereunder;
(aa) The
Company and its subsidiaries (i) are in compliance with any and all applicable foreign, federal, state and local laws and regulations
relating to the protection of human health and safety, the environment or hazardous or toxic substances or wastes, pollutants or contaminants
(“Environmental Laws”), (ii) have received all permits, licenses or other approvals required of them under applicable
Environmental Laws to conduct their respective businesses and (iii) are in compliance with all terms and conditions of such permit,
license or approval, except where such noncompliance with Environmental Laws, failure to receive required permits, licenses or other approvals
or failure to comply with the terms and conditions of such permits, licenses or approvals would not, individually or in the aggregate,
reasonably be expected to have a Material Adverse Effect;
(bb) There
are no costs or liabilities associated with Environmental Laws (including, without limitation, any capital or operating expenditures required
for clean-up, closure of properties or compliance with Environmental Laws or any permit, license or approval, any related constraints
on operating activities and any potential liabilities to third parties) which would, individually or in the aggregate, reasonably be expected
to have a Material Adverse Effect;
(cc) There
are no contracts, agreements or understandings between the Company and any person granting such person the right to require the Company
to file a registration statement under the Act with respect to any securities of the Company or to require the Company to include such
securities with the Shares registered pursuant to the Registration Statement, except as otherwise have been validly waived in connection
with the issuance and sale of Shares contemplated hereby and as described in the Pricing Disclosure Package and Prospectus;
(dd) The
Company and its subsidiaries and their respective directors, officers and employees, and to the Company’s knowledge, their respective
agents, affiliates and representatives in connection with the operation of the Company, are, and at all times have been, in compliance
with all applicable Health Care Laws (defined herein), including, but not limited to, the rules and regulations of the Food and Drug
Administration (“FDA”), the U.S. Department of Health and Human Services (“HHS”) Office of Inspector General,
the Centers for Medicare & Medicaid Services, the HHS Office for Civil Rights, the U.S. Department of Justice and any other governmental
agency or body having jurisdiction over the Company or any of its properties, and have not engaged in any activities which are, as applicable,
cause for false claims liability, civil penalties, or mandatory or permissive exclusion from Medicare, Medicaid, or any other local, state
or federal healthcare program. For purposes of this Agreement, “Health Care Laws” shall mean, to the extent applicable, the
federal Anti-kickback Statute (42 U.S.C. § 1320a-7b(b)), the Physician Payments Sunshine Act (42 U.S.C. § 1320a-7h), the civil
False Claims Act (31 U.S.C. §§ 3729 et seq.), the criminal False Claims Act (42 U.S.C. § 1320a-7b(a)), all criminal laws
relating to health care fraud and abuse, including but not limited to 18 U.S.C. Sections 286, 287 and 1349, and the health care fraud
criminal provisions under the Health Insurance Portability and Accountability Act of 1996 (42 U.S.C. §§ 1320d et seq.) (“HIPAA”),
the exclusion law (42 U.S.C. § 1320a-7), the civil monetary penalties law (42 U.S.C. § 1320a-7a), HIPAA, as amended by the Health
Information Technology for Economic and Clinical Health Act (42 U.S.C. §§ 17921 et seq.), the Federal Food, Drug, and Cosmetic
Act (21 U.S.C. §§ 301 et seq.), the Public Health Service Act (42 U.S.C. §§ 201 et seq.), the statutes, regulations
and directives of applicable federal healthcare programs, including but not limited to Medicare (Title XVIII of the Social Security Act)
and Medicaid (Title XIX of the Social Security Act), any rules and regulations promulgated pursuant to the statutes listed herein,
and any other applicable federal, state, or foreign health care laws. The Company is not a party to nor has any ongoing reporting obligations
pursuant to any corporate integrity agreement, deferred prosecution agreement, monitoring agreement, consent decree, settlement order,
plan of correction or similar agreement imposed by any governmental authority. The Company has not received any notification, correspondence
or any other written communication, including, without limitation, any FDA Form 483, notice of adverse finding, warning letter, or
untitled letter from the FDA or any similar regulatory authority, or any written notification of any pending or threatened claim, suit,
proceeding, hearing, enforcement, investigation, arbitration or other action, from any court, arbitrator, governmental or regulatory authority
or third party, of potential or actual non-compliance by, or liability of, the Company under any Health Care Laws;
(ee) Each
of the Company and its subsidiaries possesses, and is in material compliance with the terms of, all applications, certificates, approvals,
clearances, registrations, exemptions, franchises, licenses, permits, consents and other authorizations necessary to conduct their respective
businesses (collectively, “Licenses”), issued by the appropriate governmental authorities, including, without limitation,
all Licenses required by the FDA, or any component thereof and/or by any other U.S., state, local or foreign government or drug regulatory
agency (collectively, the “Regulatory Agencies”). All Licenses are in full force and effect and the Company is not in violation
of any term or conditions of any License. Each of the Company and its subsidiaries has fulfilled and performed all of its respective obligations
with respect to the Licenses and, no event has occurred which allows, or after notice or lapse of time would allow, revocation or termination
thereof or results in any other impairment of the rights of the holder of any License. The Company has not received any written notice
of proceedings relating to the revocation or modification of any Licenses and no Regulatory Agency has taken any action to limit, suspend
or revoke any License possessed by the Company;
(ff) The
pre-clinical studies and clinical trials that are described in the Registration Statement, the Pricing Prospectus and the Prospectus were
and, if still pending, are being, conducted in all material respects in accordance with all applicable laws and regulations of the Regulatory
Agencies overseeing such studies and trials; the descriptions of the pre-clinical studies and clinical trials conducted by or, to the
Company’s knowledge, on behalf of the Company, and the results thereof, contained in the Registration Statement, the Pricing Prospectus
and the Prospectus are accurate in all material respects; the Company is not aware of any other pre-clinical studies or clinical trials,
the results of which reasonably call into question the results described in the Registration Statement, the Pricing Prospectus and the
Prospectus; and the Company has not received any written notices or correspondence from the FDA, any foreign, state or local governmental
body exercising comparable authority or any Institutional Review Board requiring the termination, suspension, material modification or
clinical hold of any pre-clinical studies or clinical trials conducted by or on behalf of the Company;
(gg) Neither
the Company nor its subsidiaries nor, to the Company’s knowledge, any of its or their respective officers, employees, directors,
agents or clinical investigators, has been excluded, suspended or debarred from participation in any U.S. federal health care program
or human clinical research or, is subject to a governmental inquiry, investigation, proceeding, or other similar action that could reasonably
be expected to result in debarment, suspension, or exclusion, or has been convicted of any crime or engaged in any conduct that would
reasonably be expected to result in debarment under 42 U.S.C. § 1320a-7 or 21 U.S.C. § 335a;
(hh) To
the Company’s knowledge, the Company and its subsidiaries own, or possess valid and enforceable license rights in, all patents,
patent applications, inventions, copyrights and registrations and applications thereof, works of authorship, know how (including trade
secrets and other unpatented and/or unpatentable proprietary or confidential information, systems or procedures), trademarks, service
marks, domain names, trade names, licenses, software, systems and technology (collectively, “Intellectual Property”) necessary
for or used in the conduct or the proposed conduct of their respective businesses in the manner described in the Registration Statement,
the Pricing Prospectus and the Prospectus (“Company Intellectual Property”), provided that nothing in this sentence shall
be construed as a representation or warranty regarding the non-infringement or non-misappropriation of any Intellectual Property owned
by any third party. To the Company’s knowledge, the Company and its subsidiaries, and the conduct of their respective businesses
and the proposed conduct of their businesses, does not and will not infringe, misappropriate or otherwise violate any Intellectual Property
rights of others that would reasonably be expected to have a Material Adverse Effect. Neither the Company nor any of its subsidiaries
has received any written notice of infringement of or conflict with asserted rights of others with respect to any of the Company Intellectual
Property and there is no pending or to the Company’s knowledge, threatened action, suit, proceeding or claim by others (i) challenging
the Company’s or its subsidiaries’ rights in or to any Company Intellectual Property, and the Company and its subsidiaries
are unaware of any facts which would form a reasonable basis for any such action, suit, proceeding or claim; (ii) challenging the
validity, enforceability or scope of any Company Intellectual Property, and the Company and its subsidiaries are unaware of any facts
which would form a reasonable basis for any such action, suit, proceeding or claim; or (iii) asserting that the Company or any of
its subsidiaries infringes or otherwise violates, or would, upon the commercialization of any product or service described in the Pricing
Prospectus or the Prospectus as under development, infringe or violate, any Intellectual Property of third parties, and the Company and
its subsidiaries have not received any written notice of, and are unaware of any facts which would form a reasonable basis for any such
action, suit, proceeding or claim. To the Company’s knowledge, the Company Intellectual Property is enforceable, subsisting and,
to the knowledge of Company, valid. To the Company’s knowledge, the Company Intellectual Property is free and clear of all material
liens, security interests, or encumbrances, other than non-exclusive and exclusive licenses granted in the ordinary course of business.
To the Company’s knowledge, there are no material defects in any of the patents or patent applications, trademark applications or
trademark registration, or copyright registrations included in the Company Intellectual Property. To the Company’s knowledge, there
is no infringement, misappropriation or other violation by third parties of any Company Intellectual Property owned by the Company or
its subsidiaries. To the Company’s knowledge, the Company and its subsidiaries have complied with the terms of each agreement pursuant
to which Company Intellectual Property has been licensed to the Company or any subsidiary, and all such agreements are in full force and
effect, except as would not have a Material Adverse Effect. To the Company’s knowledge, the Company and its subsidiaries have taken
all reasonable steps to protect, maintain and safeguard the Company Intellectual Property, including payment of applicable maintenance
fees, filing of applicable statements of use, timely responding to office actions, complying with the duties of candor and good faith
required by the United States Patent and Trademark Office, and the disclosure of any required information, and the execution of appropriate
nondisclosure, confidentiality agreements, invention or intellectual property assignments or assignment agreements with their employees,
consultants and contractors. The Company has taken all reasonable steps necessary to secure assignments to its title, rights and interests
in and to the Company Intellectual Property from its employees, consultants, agents, and contractors. To the Company’s knowledge,
no employee, consultant or independent contractor of the Company is in or has ever been in violation in any material respect of any term
of any invention assignment agreement, non-competition agreement, non-solicitation agreement, or nondisclosure agreement. The Company
and each of its subsidiaries have taken reasonable measures to protect their confidential information and trade secrets, including the
execution of appropriate nondisclosure and confidentiality agreements. To the Company’s knowledge, no technology employed by the
Company or its subsidiaries has been obtained or is being used by the Company or its subsidiaries in violation of any contractual or legal
obligation binding on the Company, its subsidiaries, or any of their officers, directors, employees, or contractors, which violation relates
to the breach of a confidentiality obligation, an obligation to assign Intellectual Property to a previous employer, or an obligation
otherwise not to use the Intellectual Property of any third party. To the Company’s knowledge, there is no software licensed under
an “open source” or similar licensing model that meets the definition of “open source” promulgated by the Open
Source Initiative located online at http://opensource.org/osd (e.g., GNU General Public License, GNU Lesser General Public License, and
GNU Affero General Public License) that is licensed by the Company or its subsidiaries that is being used by the Company or its subsidiaries:
(1) in violation of any license terms applicable to the Company’s or any of its subsidiaries’ use of such open source
software; or (2) in a manner that requires any software owned by the Company or any of its subsidiaries to be made available in source
code form or be redistributable for no license fee;
(ii) No
labor dispute with the employees of the Company or any of its subsidiaries exists or, to the knowledge of the Company, is imminent, and
the Company is not aware of any existing, threatened or imminent labor disturbance by the employees of any of its principal suppliers
or contractors, except in each case as would not have a Material Adverse Effect;
(jj) (A) The
Company and its subsidiaries, taken as a whole, are insured by insurers of recognized financial responsibility against such losses and
risks and in such amounts as are prudent and customary in the businesses in which they are engaged; (B) neither the Company nor any
of its subsidiaries has been refused any insurance coverage sought or applied for; and (C) neither the Company nor any of its subsidiaries
has any reason to believe that it will not be able to renew its existing insurance coverage as and when such coverage expires or to obtain
similar coverage from similar insurers as may be necessary to continue its business at a cost that would not, taken as a whole, be reasonably
expected to have a Material Adverse Effect;
(kk) Except
as otherwise disclosed in the Registration Statement, the Pricing Prospectus and the Prospectus, the Company has not sold, issued or distributed
any shares of its common stock during the six-month period preceding the date of this Agreement, including any sales pursuant to Regulation
D or S of the Act, other than shares issued pursuant to employee benefit plans, qualified stock option plans or other employee compensation
plans or pursuant to outstanding options, rights or warrants;
(ll) All
U.S. federal income tax returns of the Company and its subsidiaries required by law to be filed have been filed and all taxes shown as
due on such returns or that otherwise have been assessed that are due and payable have been paid, except assessments against which appeals
have been promptly taken and as to which adequate reserves have been established in the audited financial statements included in the Pricing
Prospectus. The Company and each of its subsidiaries have timely filed all other U.S. federal, state, local and foreign tax returns required
to be filed through the date of this Agreement, or have requested extensions thereof, and have paid all taxes required to be paid thereon
(except where the failure to file or pay would not, individually or in the aggregate, have a Material Adverse Effect, or except as currently
being contested in good faith and for which adequate reserves have been established in the audited financial statements included in the
Pricing Prospectus), no tax deficiency has been determined adversely to the Company or any of its subsidiaries which has had (nor does
the Company nor any of its subsidiaries have any notice or knowledge of any tax deficiency that could reasonably be expected to be determined
adversely to the Company or its subsidiaries and that could reasonably be expected to have) a Material Adverse Effect and no audit or
other examination of or with respect to any tax return or taxes of the Company or any of its subsidiaries is presently in progress, nor
has the Company or any of its subsidiaries been notified in writing of any request for an audit or other examination that has not been
finally resolved and that in each case could, individually or in the aggregate, reasonably be expected to have a Material Adverse Effect;
(mm) From
the time of initial confidential submission of a registration statement relating to the Shares with the Commission through the date of
this Agreement, the Company has been and is an “emerging growth company” as defined in Section 2(a)(19) of the Act (an
“Emerging Growth Company”);
(nn) The
Company’s and its subsidiaries’ information technology assets and equipment, computers, systems, networks, hardware, software,
websites, applications and databases, and any other information technology systems owned, licensed, leased, or otherwise used by the Company,
and any such equipment or technology maintained or provided by any third parties to the Company or its subsidiaries (collectively, “IT
Systems”) are (A) adequate for, and operate and perform in all material respects as necessary for the uninterrupted operation
of the business of the Company and its subsidiaries as currently conducted and as currently planned to be conducted as described in the
Registration Statement, the Pricing Disclosure Package, and the Prospectus, and (B) free and clear of all material bugs, errors,
defects, Trojan horses, time bombs, malware, ransomware, harmful code, or other corruptants. The Company and its subsidiaries have at
all times implemented, maintained, and complied in material respects with reasonable and appropriate controls, policies, procedures and
safeguards to maintain and protect the privacy, confidentiality, integrity, availability, continuous operation, redundancy and security
of all IT Systems and all (A) information that (i) relates to, identifies, describes, or is reasonably capable of being associated
with, an identified or identifiable individual, or (ii) is defined as “personal information”, “personally identifiable
information”, “personal data”, or similar term under applicable Data Protection Requirements (as defined below) (collectively,
“Personal Data”), and (B) confidential, sensitive, or proprietary data (collectively (A) and (B) comprising
“Protected Information”)) used in connection with the business of the Company or its subsidiaries. The Company and its subsidiaries
have at all times established, maintained, and complied with reasonable and appropriate disaster recovery and security plans and procedures
for their business, including, without limitation, for the IT Systems and Protected Information. The security programs of the Company
and its subsidiaries comply in material respects with and have at all times complied in material respects, with all applicable Data Protection
Requirements. There have been no breaches or other cyberattacks involving the material accidental, unlawful, or unauthorized destruction
of, loss of, alteration of, processing of, uses of, or accesses to, any IT Systems or Protected Information, including any that have resulted
in an obligation under applicable Data Protection Requirements for the Company to notify a governmental or regulatory authority (collectively,
a “Data Breach”). There are no actual or reasonably suspected Data Breaches currently under internal review or investigations
relating to the same. The Company and its subsidiaries have not been notified of, and have no knowledge of, any fact or circumstance that
would reasonably be expected to result in any such Data Breach.
(oo) The
Company and its subsidiaries comply in all material respects and have at all times complied in all material respects with all of the following,
as they relate to (i) the use of artificial intelligence/machine learning tools, technology, and/or products and (ii) the protection,
collection, use, disclosure, transfer, storage, disposal, confidentiality, integrity, availability, processing, privacy, and security
of IT Systems and Protected Information: (A) applicable laws, statutes, regulations, rules, directives, self-regulatory guidelines
and standards, in all relevant jurisdictions; (B) all judgments, orders, rules and regulations of any court or arbitrator or
governmental or regulatory authority; (C) published policies; and (D) contractual obligations (collectively, “Data Protection
Requirements”).
(pp) The
Company and its subsidiaries have at all times made all disclosures to, and obtained all consents from, individuals as required by Data
Protection Requirements for the Company’s and its subsidiaries’ collection, use, disclosure or other processing of Protected
Information. None of such disclosures have been inaccurate, deceptive, misleading, incomplete, or in violation of Data Protection Requirements
in any material respect.
(qq) Neither
the Company nor any of its subsidiaries (A) have received any written claim or notice of any actual or potential liability under
or relating to, or actual or potential violation of, Data Protection Requirements, (B) have any knowledge of any fact or circumstance
that would reasonably indicate the Company or one of its subsidiaries is not in compliance with any of the Data Protection Requirements;
(C) are not currently participating, in whole or in part, in any investigation, remediation, or other corrective action relating
to a violation of any of the Data Protection Requirements; and (D) are not a party to any order, decree, or agreement by any court
or arbitrator or governmental or regulatory authority that imposes any obligation or liability relating to any Data Protection Requirement.
(rr) Nothing
has come to the attention of the Company that has caused the Company to believe that the statistical and market-related data included
in each of the Registration Statement, the Pricing Prospectus, the Prospectus, each Issuer Free Writing Prospectus and each Section 5(d) Writing,
as supplemented by and taken together with the Pricing Disclosure Package, is not based on or derived from sources that are reliable and
accurate in all material respects;
(ss) There
is and has been no failure on the part of the Company or any of the Company’s directors or officers, in their capacities as such,
to comply with any applicable provision of the Sarbanes-Oxley Act of 2002, as amended and the rules and regulations promulgated in
connection therewith (the “Sarbanes-Oxley Act”), including Section 402 related to loans and Sections 302 and 906 related
to certifications;
(tt) There
are no debt securities or preferred stock issued or guaranteed by the Company that are rated by a “nationally recognized statistical
rating organization,” as such term is defined under Section 3(a)(62) under the Exchange Act;
(uu) (i) Each
employee benefit plan, within the meaning of Section 3(3) of the Employee Retirement Income Security Act of 1974, as amended
(“ERISA”), for which the Company or any member of its “Controlled Group” (defined as any entity, whether or not
incorporated, that is under common control with the Company within the meaning of Section 4001(a)(14) of ERISA or any entity that
would be regarded as a single employer with the Company under Section 414(b),(c),(m) or (o) of the IRC) would have any
liability (each, a “Plan”) has been maintained in compliance with its terms and the requirements of any applicable statutes,
orders, rules and regulations, including but not limited to ERISA and the IRC; (ii) no prohibited transaction, within the meaning
of Section 406 of ERISA or Section 4975 of the IRC, has occurred with respect to any Plan, excluding transactions effected pursuant
to a statutory or administrative exemption; (iii) for each Plan that is subject to the funding rules of Section 412 of
the IRC or Section 302 of ERISA, no Plan has failed (whether or not waived), or is reasonably expected to fail, to satisfy the minimum
funding standards (within the meaning of Section 302 of ERISA or Section 412 of the IRC) applicable to such Plan; (iv) no
Plan is, or is reasonably expected to be, in “at risk status” (within the meaning of Section 303(i) of ERISA) and
no Plan that is a “multiemployer plan” within the meaning of Section 4001(a)(3) of ERISA is in “endangered
status” or “critical status” (within the meaning of Sections 304 and 305 of ERISA); (v) no “reportable event”
(within the meaning of Section 4043(c) of ERISA and the regulations promulgated thereunder) has occurred or is reasonably expected
to occur; (vi) each Plan that is intended to be qualified under Section 401(a) of the IRC is so qualified, and nothing
has occurred, whether by action or by failure to act, which would cause the loss of such qualification; (vii) neither the Company
nor any member of the Controlled Group has incurred, nor reasonably expects to incur, any liability under Title IV of ERISA (other than
contributions to the Plan or premiums to the Pension Benefit Guaranty Corporation, in the ordinary course and without default) in respect
of a Plan (including a “multiemployer plan” within the meaning of Section 4001(a)(3) of ERISA); and (viii) none
of the following events has occurred or is reasonably likely to occur: (A) a material increase in the aggregate amount of contributions
required to be made to all Plans by the Company or its Controlled Group affiliates in the current fiscal year of the Company and its Controlled
Group affiliates compared to the amount of such contributions made in the Company’s and its Controlled Group affiliates’ most
recently completed fiscal year; or (B) a material increase in the Company and its subsidiaries’ “accumulated post-retirement
benefit obligations” (within the meaning of Accounting Standards Codification Topic 715-60) compared to the amount of such obligations
in the Company and its subsidiaries’ most recently completed fiscal year, except in each case with respect to the events or conditions
set forth in (i) through (viii) hereof, as would not, individually or in the aggregate, have a Material Adverse Effect;
(vv) There
are no contracts, arrangements or other documents that are required to be described in the Registration Statement, the Pricing Prospectus
or the Prospectus or to be filed as exhibits to the Registration Statement that are not described or filed as required;
(ww) The
Company and each of its subsidiaries have such permits, licenses, approvals, consents, franchises, certificates of need and other approvals
or authorizations of governmental or regulatory authorities (“Permits”) as are necessary under applicable law to own their
respective properties and conduct their respective businesses in the manner described in the Registration Statement, the Pricing Prospectus
and the Prospectus, except for any of the foregoing that would not, individually or in the aggregate, have a Material Adverse Effect.
Neither the Company nor any of its subsidiaries has received notice of any proceedings related to the revocation or modification of any
such Permits that, individually or in the aggregate, if the subject of an unfavorable decision, ruling or finding, would have a Material
Adverse Effect;
(xx) Neither
the Company nor any of its subsidiaries or controlled affiliates has taken or will take, directly or indirectly, any action designed to
or that could reasonably be expected to cause or result in the stabilization or manipulation of the price of any security of the Company
or any of its subsidiaries in connection with the offering of the Shares;
(yy) The
Company is not a party to any contract, agreement or understanding with any person (other than this Agreement) that would give rise to
a valid claim against the Company or any Underwriter for a brokerage commission, finder’s fee or like payment in connection with
the offering and sale of the Shares;
(zz) No
relationship, direct or indirect, exists between or among the Company on the one hand, and the directors, officers, stockholders, customers
or suppliers of the Company, on the other, that is required by the Act to be described in the Registration Statement and the Prospectus
and that is not so described in such documents and in the Registration Statement, the Pricing Prospectus and the Prospectus;
(aaa) No
forward-looking statement (within the meaning of Section 27A of the Act and Section 21E of the Exchange Act) included in any
of the Registration Statement, the Pricing Prospectus or the Prospectus has been made or reaffirmed without a reasonable basis or has
been disclosed other than in good faith.
(bbb) Neither
the Company nor any of its subsidiaries is a “covered foreign person”, as that term is defined in 31 C.F.R. § 850.209.
Neither the Company nor any of its subsidiaries currently engages, or has plans to engage, directly or indirectly, in a “covered
activity”, as that term is defined in in 31 C.F.R. § 850.208 (“Covered Activity”). The Company does not have any
joint ventures that engage in or plan to engage in any Covered Activity. The Company also does not, directly or indirectly, hold a board
seat on, have a voting or equity interest in, or have any contractual power to direct or cause the direction of the management or policies
of any person or persons that engages or plans to engage in any Covered Activity.
2. Subject
to the terms and conditions herein set forth, (a) the Company agrees to issue and sell to each of the Underwriters, and each of the
Underwriters agrees, severally and not jointly, to purchase from the Company, at a purchase price per share of $[·], the number
of Firm Shares set forth opposite the name of such Underwriter in Schedule I hereto and (b) in the event and to the extent that the
Underwriters shall exercise the election to purchase Optional Shares as provided below, the Company agrees to issue and sell to each of
the Underwriters, and each of the Underwriters agrees, severally and not jointly, to purchase from the Company, at the purchase price
per share set forth in clause (a) of this Section 2 (provided that the purchase price per Optional Share shall be reduced by
an amount per share equal to any dividends or distributions declared by the Company and payable on the Firm Shares but not payable on
the Optional Shares), that portion of the number of Optional Shares as to which such election shall have been exercised (to be adjusted
by you so as to eliminate fractional shares) determined by multiplying such number of Optional Shares by a fraction, the numerator of
which is the maximum number of Optional Shares which such Underwriter is entitled to purchase as set forth opposite the name of such Underwriter
in Schedule I hereto and the denominator of which is the maximum number of Optional Shares that all of the Underwriters are entitled to
purchase hereunder.
The Company hereby grants
to the Underwriters the right to purchase at their election up to [•] Optional Shares, at the purchase price per share set forth
in the paragraph above, provided that the purchase price per Optional Share shall be reduced by an amount per share equal to any dividends
or distributions declared by the Company and payable on the Firm Shares but not payable on the Optional Shares. Any such election to purchase
Optional Shares may be exercised only by written notice from you to the Company, given within a period of 30 calendar days after the date
of this Agreement, setting forth the aggregate number of Optional Shares to be purchased and the date on which such Optional Shares are
to be delivered, as determined by you but in no event earlier than the First Time of Delivery (as defined in Section 4 hereof) or,
unless you and the Company otherwise agree in writing, earlier than one or later than ten business days after the date of such notice.
3. Upon
the authorization by you of the release of the Shares, the several Underwriters propose to offer the Shares for sale upon the terms and
conditions set forth in the Pricing Disclosure Package and the Prospectus.
4. (a) The
Shares to be purchased by each Underwriter hereunder, in definitive or book-entry form, and in such authorized denominations and registered
in such names as the Representatives may request upon at least twenty-four hours’ prior notice to the Company shall be delivered
by or on behalf of the Company to the Representatives, through the facilities of the Depository Trust Company (“DTC”), for
the account of such Underwriter, against payment by or on behalf of such Underwriter of the purchase price therefor by wire transfer of
Federal (same-day) funds to the account specified by the Company to the Representatives at least twenty-four hours in advance. The Company
will cause the certificates, if any, representing the Shares to be made available for checking and packaging at least twenty-four hours
prior to the Time of Delivery (as defined below) with respect thereto at the office of DTC or its designated custodian (the “Designated
Office”). The time and date of such delivery and payment shall be, with respect to the Firm Shares, 9:30 a.m., New York time, on
[•], 2026 or such other time and date as the Representatives and the Company may agree upon in writing, and, with respect to the
Optional Shares, 9:30 a.m., New York time, on the date specified by the Representatives in the written notice given by the Representatives
of the Underwriters’ election to purchase such Optional Shares, or such other time and date as the Representatives and the Company
may agree upon in writing. Such time and date for delivery of the Firm Shares is herein called the “First Time of Delivery”,
such time and date for delivery of the Optional Shares, if not the First Time of Delivery, is herein called the “Second Time of
Delivery”, and each such time and date for delivery is herein called a “Time of Delivery.”
(b) The
documents to be delivered at each Time of Delivery by or on behalf of the parties hereto pursuant to Section 8 hereof, including
the cross receipt for the Shares and any additional documents requested by the Underwriters pursuant to Section 8(l) hereof,
will be delivered at the offices of Cooley LLP, 55 Hudson Yards, New York, New York 10001 (the “Closing Location”), and the
Shares will be delivered at the Designated Office, all at such Time of Delivery. A meeting will be held at the Closing Location at [•]
p.m., New York time, on the New York Business Day next preceding such Time of Delivery, at which meeting the final drafts of the documents
to be delivered pursuant to the preceding sentence will be available for review by the parties hereto. For the purposes of this Section 4,
“New York Business Day” shall mean each Monday, Tuesday, Wednesday, Thursday and Friday which is not a day on which banking
institutions in New York City are generally authorized or obligated by law or executive order to close.
5. The
Company agrees with each of the Underwriters:
(a) To
prepare the Prospectus in a form approved by you and to file such Prospectus pursuant to Rule 424(b) under the Act prior to
the earlier of (i) the First Time of Delivery and (ii) the Commission’s close of business on the second business day following
the execution and delivery of this Agreement, or, if applicable, such earlier time as may be required by Rule 430A(a)(3) under
the Act; to make no further amendment or any supplement to the Registration Statement or the Prospectus prior to the last Time of Delivery
which shall be disapproved by you promptly after reasonable notice thereof; to advise you, promptly after it receives notice thereof,
of the time when any amendment to the Registration Statement has been filed or becomes effective or any amendment or supplement to the
Prospectus has been filed and to furnish you with copies thereof; to file promptly all material required to be filed by the Company with
the Commission pursuant to Rule 433(d) under the Act; to advise you, promptly after it receives notice thereof, of the issuance
by the Commission of any stop order or of any order suspending the effectiveness of the Registration Statement or any part thereof preventing
or suspending the use of any Issuer Free Writing Prospectus, Preliminary Prospectus or other prospectus in respect of the Shares, of the
suspension of the qualification of the Shares for offering or sale in any jurisdiction, of the initiation or threatening of any proceeding
for any such purpose or pursuant to Section 8A of the Act, or of any request by the Commission for the amending or supplementing
of the Registration Statement or the Prospectus or for additional information; and, in the event of the issuance of any stop order suspending
the effectiveness of the Registration Statement or any part thereof or of any order preventing or suspending the use of any Issuer Free
Writing Prospectus, Preliminary Prospectus or other prospectus relating to the Shares or suspending any such qualification, to promptly
use its best efforts to obtain the withdrawal of such order;
(b) Promptly
from time to time to take such action as you may reasonably request to qualify the Shares for offering and sale under the applicable securities
laws of such jurisdictions as you may request and to comply with such laws so as to permit the continuance of sales and dealings therein
in such jurisdictions for as long as may be necessary to complete the distribution of the Shares, provided that in connection therewith
the Company shall not be required to qualify as a foreign corporation (where not otherwise required), or to file a general consent to
service of process in any jurisdiction (where not otherwise required);
(c) Prior
to 10:00 a.m., New York time, on the New York Business Day next succeeding the date of this Agreement (or such other time as may be agreed
to by the Representatives and the Company) and from time to time, to furnish the Underwriters with written and electronic copies of the
Prospectus, Preliminary Prospectus and any supplements and amendments thereto or to the Registration Statement in such quantities as the
Representatives may reasonably request, and, if the delivery of a prospectus (or in lieu thereof, the notice referred to in Rule 173(a) under
the Act) is required at any time prior to the expiration of nine months after the time of issue of the Prospectus in connection with the
offering or sale of the Shares and if at such time any event shall have occurred as a result of which the Prospectus as then amended or
supplemented would include an untrue statement of a material fact or omit to state any material fact necessary in order to make the statements
therein, in the light of the circumstances under which they were made when such Prospectus (or in lieu thereof, the notice referred to
in Rule 173(a) under the Act) is delivered, not misleading, or, if for any other reason it shall be necessary during such same
period to amend or supplement the Prospectus in order to comply with the Act, to notify you and, before amending or supplementing the
Registration Statement, the Pricing Disclosure Package or the Prospectus, to furnish you a copy of each such proposed amendment or supplement
and not file any such proposed amendment or supplement to which you reasonably object, and upon your request to prepare and furnish without
charge to each Underwriter and to any dealer in securities (whose name and address the Underwriters shall furnish to the Company) as many
written and electronic copies as you may from time to time reasonably request of an amended Prospectus or a supplement to the Prospectus
which will correct such statement or omission or effect such compliance; and in case any Underwriter is required to deliver a prospectus
(or in lieu thereof, the notice referred to in Rule 173(a) under the Act) in connection with sales of any of the Shares at any
time nine months or more after the time of issue of the Prospectus, upon your request but at the expense of such Underwriter, to prepare
and deliver to such Underwriter as many written and electronic copies as you may request of an amended or supplemented Prospectus complying
with Section 10(a)(3) of the Act;
(d) To
make generally available to its securityholders as soon as practicable (which may be satisfied by filing with the Commission’s Electronic
Data Gathering, Analysis and Retrieval System (“EDGAR”)), but in any event not later than sixteen months after the effective
date of the Registration Statement (as defined in Rule 158(c) under the Act), an earnings statement of the Company and its subsidiaries
(which need not be audited) complying with Section 11(a) of the Act and the rules and regulations of the Commission thereunder
(including, at the option of the Company, Rule 158);
(e) (1) During
the period beginning from the date of this Agreement and continuing to and including the date 180 days after the date of the Prospectus
(the “Lock-Up Period”), not to (i) offer, sell, contract to sell, pledge, grant any option to purchase, loan, hedge,
make any short sale or otherwise transfer or dispose of, directly or indirectly, or file with or confidentially submit to the Commission
a registration statement under the Act relating to, any securities of the Company that are substantially similar to the Shares, including
but not limited to any options or warrants to purchase shares of Stock or any securities that are convertible into or exchangeable for,
or that represent the right to receive, Stock or any such substantially similar securities, (ii) enter into any hedging, swap or
other agreement that transfers, in whole or in part, any of the economic consequences of ownership of the Stock or any such other securities,
whether any such transaction described in clause (i) or (ii) above is to be settled by delivery of Stock or such other securities,
in cash or otherwise (other than the Shares to be sold hereunder or pursuant to employee stock option plans existing on, or upon the conversion
or exchange of convertible or exchangeable securities outstanding as of, the date of this Agreement) or (iii) publicly disclose the
intention to do any of the foregoing, in each case, without the prior written consent of Goldman Sachs & Co. LLC and Jefferies
LLC; provided, however, that the restrictions in the foregoing sentence shall not apply to (a) the Shares to be sold hereunder; (b) shares
of Stock or any securities convertible into, or exercisable for, shares of Stock issued by the Company upon exercise or settlement of
options, warrants or restricted stock units outstanding as of the date hereof and described in the Pricing Disclosure Package; (c) the
grant of any shares of Stock or any securities convertible into, or exercisable for, shares of Stock (including without limitation options,
warrants, restricted stock or restricted stock units) pursuant to employee equity-based compensation plans, incentive plans, stock plans,
dividend reinvestment plans or other arrangements in effect as of the date hereof and described in the Pricing Disclosure Package; (d) the
filing of a registration statement on Form S-8 in connection with the registration of securities issuable under any employee equity-based
compensation plan, incentive plan, stock plan, or dividend reinvestment plan adopted and approved by the Company’s board of directors
prior to the date hereof and described in the Pricing Disclosure Package; or (e) shares of Stock or other securities issued in connection
with any bona fide merger, joint venture, strategic alliance, commercial or other collaborative transaction, or the acquisition or license
by the Company of the business, property, technology or other assets of another individual or entity that is an unaffiliated third party
of the Company, or the assumption of an employee benefit plan in connection with such a merger or acquisition, provided that (i) the
aggregate number of shares issued pursuant to this clause (e) (on an as-converted or as-exercised basis, as the case may be) shall
not exceed 5% of the total number of outstanding shares of Stock immediately following the issuance and sale of the Shares hereunder and
(ii) the recipient of any such shares of Stock or securities issued pursuant to this clause (e) during such period shall enter
into an agreement substantially in the form of Annex II hereto;
(2) If
Goldman Sachs & Co. LLC and Jefferies LLC, in their sole discretion, agree to release or waive the restrictions in lock-up letters
delivered pursuant to Section 8(i) hereof, in each case, for an officer or director of the Company and provide the Company with
notice of the impending release or waiver at least three business days before the effective date of the release or waiver, the Company
agrees to announce the impending release or waiver by a press release substantially in the form of Annex I hereto through a major news
service at least two business days before the effective date of the release or waiver.
(f) During
a period of three years from the effective date of the Registration Statement, so long as the Company is subject to the reporting requirements
of either Section 13 or Section 15(d) of the Exchange Act, to furnish to its stockholders as soon as practicable after
the end of each fiscal year an annual report (including a balance sheet and statements of operations and comprehensive loss, convertible
preferred stock and stockholders' deficit and cash flows of the Company and its consolidated subsidiaries certified by independent public
accountants) and, as soon as practicable after the end of each of the first three quarters of each fiscal year (beginning with the fiscal
quarter ending after the effective date of the Registration Statement), to make available to its stockholders consolidated summary financial
information of the Company and its subsidiaries for such quarter in reasonable detail; provided that no reports, documents or other information
need to be furnished pursuant to this Section 5(f) to the extent that they are available on the Commission’s EDGAR system;
(g) During
a period of three years from the effective date of the Registration Statement, so long as the Company is subject to the reporting requirements
of either Section 13 or Section 15(d) of the Exchange Act, to furnish to you copies of all reports or other communications
(financial or other) furnished to stockholders, and to deliver to you (i) as soon as they are available, copies of any reports and
financial statements furnished to or filed with the Commission or any national securities exchange on which any class of securities of
the Company is listed; and (ii) such additional information concerning the business and financial condition of the Company as you
may from time to time reasonably request (such financial statements to be on a consolidated basis to the extent the accounts of the Company
and its subsidiaries are consolidated in reports furnished to its stockholders generally or to the Commission); provided that no reports,
documents or other information need to be furnished pursuant to this Section 5(g) to the extent that they are available on the
Commission’s EDGAR system;
(h) To
use the net proceeds received by it from the sale of the Shares pursuant to this Agreement in the manner specified in the Pricing Prospectus
under the caption “Use of Proceeds”;
(i) To
use its best efforts to list for trading, subject to official notice of issuance, the Shares on the Exchange;
(j) To
file with the Commission such information on Form 10-Q or Form 10-K as may be required by Rule 463 under the Act;
(k) If
the Company elects to rely upon Rule 462(b) under the Act, the Company shall file a Rule 462(b) Registration Statement
with the Commission in compliance with Rule 462(b) under the Act by 10:00 P.M., Washington, D.C. time, on the date of this Agreement,
and the Company shall at the time of filing either pay to the Commission the filing fee for the Rule 462(b) Registration Statement
or give irrevocable instructions for the payment of such fee pursuant to Rule 111(b) under the Act;
(l) Upon
reasonable request of any Underwriter, to furnish, or cause to be furnished, to such Underwriter an electronic version of the Company’s
trademarks, servicemarks and corporate logo for use on the website, if any, operated by such Underwriter for the purpose of facilitating
the on-line offering of the Shares (the “License”); provided, however, that the License shall be used solely for the purpose
described above, is granted without any fee and may not be assigned or transferred;
(m) To
promptly notify you if the Company ceases to be an Emerging Growth Company at any time prior to the later of (i) completion of the
distribution of the Shares within the meaning of the Act and (ii) the last Time of Delivery.
(n) To
indemnify and hold harmless the Underwriters against any documentary, stamp, registration or other similar tax, including any interest
and penalties, on the sale, issuance or delivery of the Shares by the Company to the Underwriters and on the execution and delivery of
this Agreement; provided that the foregoing shall not apply to stock transfer taxes on resale of any of the Shares by the Underwriters.
6. (a) The
Company represents and agrees that, without the prior consent of the Representatives, it has not made and will not make any offer relating
to the Shares that would constitute a “free writing prospectus” as defined in Rule 405 under the Act; each Underwriter
represents and agrees that, without the prior consent of the Company and the Representatives, it has not made and will not make any offer
relating to the Shares that would constitute a free writing prospectus required to be filed with the Commission; and any such free writing
prospectus the use of which has been consented to by the Company and the Representatives is listed on Schedule II(a) hereto;
(b) The
Company has complied and will comply with the requirements of Rule 433 under the Act applicable to any Issuer Free Writing Prospectus,
including timely filing with the Commission or retention where required and legending; and the Company represents that it has satisfied
and agrees that it will satisfy the conditions under Rule 433 under the Act to avoid a requirement to file with the Commission any
electronic road show;
(c) The
Company agrees that if at any time following issuance of an Issuer Free Writing Prospectus or Written Testing-the-Waters Communication
any event occurred or occurs as a result of which such Issuer Free Writing Prospectus or Written Testing-the-Waters Communication would
conflict with the information in the Registration Statement, the Pricing Prospectus or the Prospectus or would include an untrue statement
of a material fact or omit to state any material fact necessary in order to make the statements therein, in the light of the circumstances
then prevailing, not misleading, the Company will give prompt notice thereof to the Representatives and, if requested by the Representatives,
will prepare and furnish without charge to each Underwriter an Issuer Free Writing Prospectus, Written Testing-the-Waters Communication
or other document which will correct such conflict, statement or omission; provided, however, that this covenant shall not apply to any
statements or omissions in a Written Testing-the-Waters Communication prepared or authorized by the Company made in reliance upon and
in conformity with the Underwriter Information;
(d) The
Company represents and agrees that (i) it has not engaged in, or authorized any other person to engage in, any Testing-the-Waters
Communications, other than Testing-the-Waters Communications with the prior consent of the Representatives with entities that the Company
reasonably believes are qualified institutional buyers as defined in Rule 144A under the Act or institutions that are accredited
investors as defined in Rule 501(a)(1), (a)(2), (a)(3), (a)(7), (a)(8), (a)(9), (a)(12) or (a)(13) under the Act, and (ii) it
has not distributed, or authorized any other person to distribute, any Written Testing-the-Waters Communication, other than those distributed
with the prior consent of the Representatives that are listed on Schedule III(d) hereto; and the Company reconfirms that the Underwriters
have been authorized to act on its behalf in engaging in Testing-the-Waters Communications; and
(e) Each
Underwriter represents and agrees that any Testing-the-Waters Communications undertaken by it were with entities that such Underwriter
reasonably believes are qualified institutional buyers as defined in Rule 144A under the Act or institutions that are accredited
investors as defined in Rule 501(a)(1), (a)(2), (a)(3), (a)(7), (a)(8), (a)(9), (a)(12) or (a)(13) under the Act.
7. The
Company covenants and agrees with the several Underwriters that the Company will pay or cause to be paid the following: (i) the fees,
disbursements and expenses of the Company's counsel and accountants in connection with the registration of the Shares under the Act and
all other expenses in connection with the preparation, printing, reproduction and filing of the Registration Statement, any Preliminary
Prospectus, any Written Testing-the-Waters Communication, any Issuer Free Writing Prospectus and the Prospectus and amendments and supplements
thereto and the mailing and delivering of copies thereof to the Underwriters and dealers; (ii) the cost of printing or producing
any agreement among Underwriters, this Agreement, the Blue Sky Memorandum, closing documents (including any compilations thereof) and
any other documents in connection with the offering, purchase, sale and delivery of the Shares; (iii) all expenses in connection
with the qualification of the Shares for offering and sale under state securities laws as provided in Section 5(b) hereof, including
the reasonable and documented fees and disbursements of counsel for the Underwriters in connection with such qualification and in connection
with the Blue Sky survey; (iv) all fees and expenses in connection with listing the Shares on the Exchange; (v) the filing fees
incident to, and the reasonable and documented fees and disbursements of counsel for the Underwriters in connection with, any required
review by FINRA of the terms of the sale of the Shares, provided, however, that the reasonable and documented fees and disbursements of
counsel for the Underwriters related to clauses (iii) and (v) shall not exceed $40,000 in the aggregate; (vi) the cost
of preparing stock certificates, if applicable; (vii) the cost and charges of any transfer agent or registrar; (viii) the costs
and expenses of the Company relating to investor presentations on any “road show” undertaken in connection with the marketing
of the offering of the Shares, including, without limitation, expenses associated with the preparation or dissemination of any electronic
road show, expenses associated with the production of road show slides and graphics, fees and expenses of any consultants engaged in connection
with the road show presentations with the prior approval of the Company, and travel and lodging expenses of the representatives and officers
of the Company and any such consultants; and (ix) all other costs and expenses incident to the performance of its obligations hereunder
which are not otherwise specifically provided for in this Section; and (x) any documentary, stamp, registration or other similar
taxes, if any, on the sale, issuance and delivery of the Shares by the Company to the Underwriters and on the execution and delivery of
this Agreement. It is understood, however, that, except as provided in this Section, and Sections 9 and 12 hereof, the Underwriters will
pay all of their own costs and expenses, including the fees of their counsel, stock transfer taxes on resale of any of the Shares by them,
and any advertising expenses connected with any offers they may make.
8. The
obligations of the Underwriters hereunder, as to the Shares to be delivered at each Time of Delivery, shall be subject, in their discretion,
to the condition that all representations and warranties and other statements of the Company herein are, at and as of the Applicable Time
and such Time of Delivery, true and correct, the condition that the Company shall have performed all of its obligations hereunder theretofore
to be performed, and the following additional conditions:
(a) The
Prospectus shall have been filed with the Commission pursuant to Rule 424(b) under the Act within the applicable time period
prescribed for such filing by the rules and regulations under the Act and in accordance with Section 5(a) hereof; all material
required to be filed by the Company pursuant to Rule 433(d) under the Act shall have been filed with the Commission within the
applicable time period prescribed for such filing by Rule 433 under the Act; if the Company has elected to rely upon Rule 462(b) under
the Act, the Rule 462(b) Registration Statement shall have become effective by 10:00 P.M., Washington, D.C. time, on the date
of this Agreement; no stop order suspending the effectiveness of the Registration Statement or any part thereof shall have been issued
and no proceeding for that purpose or pursuant to Section 8A of the Act shall have been initiated or threatened by the Commission;
no stop order suspending or preventing the use of the Pricing Prospectus, Prospectus or any Issuer Free Writing Prospectus shall have
been initiated or threatened by the Commission; and all requests for additional information on the part of the Commission shall have been
complied with to your reasonable satisfaction;
(b) Cooley
LLP, counsel for the Underwriters, shall have furnished to you such written opinion and negative assurance letter, dated such Time of
Delivery, in form and substance satisfactory to you, and such counsel shall have received such papers and information as they may reasonably
request to enable them to pass upon such matters;
(c) Goodwin
Procter LLP, counsel for the Company, shall have furnished to you their written opinion and negative assurance letter, dated such Time
of Delivery, in form and substance reasonably satisfactory to you;
(d) Morgan
Lewis & Bockius LLP and Day Pitney LLP, each an intellectual property counsel for the Company, shall have each furnished to you
their written opinion, dated such Time of Delivery, in form and substance reasonably satisfactory to you;
(e) On
the date of the Prospectus at a time prior to the execution of this Agreement, at 9:30 a.m., New York time, on the effective date of any
post-effective amendment to the Registration Statement filed subsequent to the date of this Agreement and also at each Time of Delivery,
PricewaterhouseCoopers LLP shall have furnished to you a letter or letters, dated the respective dates of delivery thereof, in form and
substance satisfactory to you;
(f) (i) Neither
the Company nor any of its subsidiaries shall have sustained since the date of the latest audited financial statements included in the
Pricing Prospectus any loss or interference with its business from fire, explosion, flood or other calamity, whether or not covered by
insurance, or from any labor dispute or court or governmental action, order or decree, otherwise than as set forth or contemplated in
the Pricing Prospectus, and (ii) since the respective dates as of which information is given in the Pricing Prospectus there shall
not have been any change in the capital stock (other than as a result of the exercise of stock options or the award of stock options or
restricted stock in the ordinary course of business pursuant to the Company’s equity plans that are described in the Pricing Prospectus)
or long-term debt of the Company or any of its subsidiaries or any change or effect, or any development involving a prospective change
or effect, in or affecting (x) the business, properties, general affairs, management, financial position, stockholders’ deficit
or results of operations of the Company and its subsidiaries, taken as a whole, except as set forth or contemplated in the Pricing Prospectus
or (y) the ability of the Company to perform its obligations under this Agreement, including the issuance and sale of the Shares,
or to consummate the transactions contemplated in the Pricing Prospectus and the Prospectus, the effect of which, in any such case described
in clause (i) or (ii), is in your judgment so material and adverse as to make it impracticable or inadvisable to proceed with the
public offering or the delivery of the Shares being delivered at such Time of Delivery on the terms and in the manner contemplated in
the Pricing Prospectus and the Prospectus;
(g) On
or after the Applicable Time there shall not have occurred any of the following: (i) a suspension or material limitation in trading
in securities generally on the New York Stock Exchange or the Exchange; (ii) a suspension or material limitation in trading in the
Company’s securities on the Exchange; (iii) a general moratorium on commercial banking activities declared by either Federal
or New York State authorities or a material disruption in commercial banking or securities settlement or clearance services in the United
States; (iv) the outbreak or escalation of hostilities involving the United States or the declaration by the United States of a national
emergency or war or (v) the occurrence of any other calamity or crisis or any change in financial, political or economic conditions
in the United States or elsewhere, if the effect of any such event specified in clause (iv) or (v) in your judgment makes it
impracticable or inadvisable to proceed with the public offering or the delivery of the Shares being delivered at such Time of Delivery
on the terms and in the manner contemplated in the Pricing Prospectus and the Prospectus;
(h) The
Shares to be sold at such Time of Delivery shall have been duly listed, subject to official notice of issuance for quotation on the Exchange;
(i) The
Company shall have obtained and delivered to the Underwriters executed copies of an agreement from each (i) officer and director
of the Company and (ii) other stockholders of the Company representing substantially all outstanding securities of the Company, substantially
to the effect set forth in Annex II hereto in form and substance satisfactory to you;
(j) The
Company shall have complied with the provisions of Section 5(c) hereof with respect to the furnishing of prospectuses on the
New York Business Day next succeeding the date of this Agreement; and
(k) The
Company shall have delivered to the Representatives on the date of the Prospectus at a time prior to the execution of this Agreement and
at such Time of Delivery a certificate of the Chief Financial Officer of the Company, in form and substance satisfactory to the Representatives;
and
(l) The
Company shall have furnished or caused to be furnished to you at such Time of Delivery certificates of officers of the Company satisfactory
to you as to the accuracy of the representations and warranties of the Company herein at and as of such Time of Delivery, as to the performance
by the Company of all of its obligations hereunder to be performed at or prior to such Time of Delivery, as to the matters set forth in
subsections (a) and (f) of this Section 8 and as to such other matters as you may reasonably request.
9. (a)
The Company will indemnify and hold harmless each Underwriter against any losses, claims, damages or liabilities, joint or several, to
which such Underwriter may become subject, under the Act or otherwise, insofar as such losses, claims, damages or liabilities (or actions
in respect thereof) arise out of or are based upon an untrue statement or alleged untrue statement of a material fact contained in the
Registration Statement, any Preliminary Prospectus, the Pricing Prospectus or the Prospectus, or any amendment or supplement thereto,
any Issuer Free Writing Prospectus, any “roadshow” as defined in Rule 433(h) under the Act (a “roadshow”),
any “issuer information” filed or required to be filed pursuant to Rule 433(d) under the Act or any Testing-the-Waters
Communication, or arise out of or are based upon the omission or alleged omission to state therein a material fact required to be stated
therein or necessary to make the statements therein not misleading, and will reimburse each Underwriter for any legal or other expenses
reasonably incurred by such Underwriter in connection with investigating or defending any such action or claim as such expenses are incurred;
provided, however, that the Company shall not be liable in any such case to the extent that any such loss, claim, damage
or liability arises out of or is based upon an untrue statement or alleged untrue statement or omission or alleged omission made in the
Registration Statement, any Preliminary Prospectus, the Pricing Prospectus or the Prospectus, or any amendment or supplement thereto,
or any Issuer Free Writing Prospectus, any road show, or any Testing-the-Waters Communication, in reliance upon and in conformity with
the Underwriter Information.
(b) Each
Underwriter, severally and not jointly, will indemnify and hold harmless the Company against any losses, claims, damages or liabilities
to which the Company may become subject, under the Act or otherwise, insofar as such losses, claims, damages or liabilities (or actions
in respect thereof) arise out of or are based upon an untrue statement or alleged untrue statement of a material fact contained in the
Registration Statement, any Preliminary Prospectus, the Pricing Prospectus or the Prospectus, or any amendment or supplement thereto,
or any Issuer Free Writing Prospectus, or any roadshow or any Testing-the-Waters Communication, or arise out of or are based upon the
omission or alleged omission to state therein a material fact required to be stated therein or necessary to make the statements therein
not misleading, in each case to the extent, but only to the extent, that such untrue statement or alleged untrue statement or omission
or alleged omission was made in the Registration Statement, any Preliminary Prospectus, the Pricing Prospectus or the Prospectus, or any
amendment or supplement thereto, or any Issuer Free Writing Prospectus, or any roadshow or any Testing-the-Waters Communication, in reliance
upon and in conformity with the Underwriter Information; and will reimburse the Company for any legal or other expenses reasonably incurred
by the Company in connection with investigating or defending any such action or claim as such expenses are incurred. As used in this Agreement
with respect to an Underwriter and an applicable document, “Underwriter Information” shall mean the written information furnished
to the Company by such Underwriter through the Representatives expressly for use therein; it being understood and agreed upon that the
only such information furnished by any Underwriter consists of the following information in the Prospectus furnished on behalf of each
Underwriter: the concession and reallowance figures appearing in the fifth paragraph under the caption “Underwriting”, and
the information contained in the fourteenth, fifteenth and sixteenth paragraphs under the caption “Underwriting”.
(c) Promptly
after receipt by an indemnified party under subsection (a) or (b) above of notice of the commencement of any action, such indemnified
party shall, if a claim in respect thereof is to be made against the indemnifying party under such subsection, notify the indemnifying
party in writing of the commencement thereof; provided that the failure to notify the indemnifying party shall not relieve it from any
liability that it may have under the preceding paragraphs of this Section 9 except to the extent that it has been materially prejudiced
(through the forfeiture of substantive rights or defenses) by such failure; and provided further that the failure to notify the indemnifying
party shall not relieve it from any liability that it may have to an indemnified party otherwise than under the preceding paragraphs of
this Section 9. In case any such action shall be brought against any indemnified party and it shall notify the indemnifying party
of the commencement thereof, the indemnifying party shall be entitled to participate therein and, to the extent that it shall wish, jointly
with any other indemnifying party similarly notified, to assume the defense thereof, with counsel reasonably satisfactory to such indemnified
party (who shall not, except with the consent of the indemnified party, be counsel to the indemnifying party), and, after notice from
the indemnifying party to such indemnified party of its election so to assume the defense thereof, the indemnifying party shall not be
liable to such indemnified party under such subsection for any reasonable and documented legal expenses of other counsel or any other
expenses, in each case subsequently incurred by such indemnified party, in connection with the defense thereof other than reasonable costs
of investigation; provided, however, that in any such proceeding, any indemnified party shall have the right to retain its own counsel,
but the fees and expenses of such counsel shall be at the expense of such indemnified party unless (i) the indemnifying party and
the indemnified person shall have mutually agreed to the contrary; (ii) the indemnifying party has failed within a reasonable time
to retain counsel reasonably satisfactory to the indemnified party; (iii) the indemnified party shall have reasonably concluded that
there may be legal defenses available to it that are different from or in addition to those available to the indemnifying party; or (iv) the
named parties in any such proceeding (including the impleaded parties) include both the indemnifying party and the indemnified party and
representation of both parties by the same counsel would be inappropriate due to actual or potential differing interests between them.
No indemnifying party shall, without the written consent of the indemnified party, effect the settlement or compromise of, or consent
to the entry of any judgment with respect to, any pending or threatened action or claim in respect of which indemnification or contribution
may be sought hereunder (whether or not the indemnified party is an actual or potential party to such action or claim) unless such settlement,
compromise or judgment (i) includes an unconditional release of the indemnified party from all liability arising out of such action
or claim and (ii) does not include a statement as to or an admission of fault, culpability or a failure to act, by or on behalf of
any indemnified party. Further, the indemnifying party shall not be liable for any settlement of any proceeding effected without its written
consent, but if settled with such consent, the indemnifying party agrees to indemnify the indemnified party from and against any loss
or liability by reason of such settlement. Notwithstanding the foregoing, if at any time an indemnified party shall have requested an
indemnifying party to reimburse the indemnified party for fees and expenses of counsel as contemplated by this Section 9(c), the
indemnifying party agrees that it shall be liable for any settlement of any proceeding effected without its written consent if (i) such
settlement is entered into more than 30 days after receipt by such indemnifying party of the aforesaid request and (ii) such indemnifying
party shall not have reimbursed the indemnified party in accordance with such request prior to the date of such settlement.
(d) If
the indemnification provided for in this Section 9 is unavailable to or insufficient to hold harmless an indemnified party under
subsection (a) or (b) above in respect of any losses, claims, damages or liabilities (or actions in respect thereof) referred
to therein, then each indemnifying party shall contribute to the amount paid or payable by such indemnified party as a result of such
losses, claims, damages or liabilities (or actions in respect thereof) in such proportion as is appropriate to reflect the relative benefits
received by the Company on the one hand and the Underwriters on the other from the offering of the Shares. If, however, the allocation
provided by the immediately preceding sentence is not permitted by applicable law, then each indemnifying party shall contribute to such
amount paid or payable by such indemnified party in such proportion as is appropriate to reflect not only such relative benefits but also
the relative fault of the Company on the one hand and the Underwriters on the other in connection with the statements or omissions which
resulted in such losses, claims, damages or liabilities (or actions in respect thereof), as well as any other relevant equitable considerations.
The relative benefits received by the Company on the one hand and the Underwriters on the other shall be deemed to be in the same proportion
as the total net proceeds from the offering (before deducting expenses) received by the Company bear to the total underwriting discounts
and commissions received by the Underwriters, in each case as set forth in the table on the cover page of the Prospectus. The relative
fault shall be determined by reference to, among other things, whether the untrue or alleged untrue statement of a material fact or the
omission or alleged omission to state a material fact relates to information supplied by the Company on the one hand or the Underwriters
on the other and the parties' relative intent, knowledge, access to information and opportunity to correct or prevent such statement or
omission. The Company and the Underwriters agree that it would not be just and equitable if contribution pursuant to this subsection (d) were
determined by pro rata allocation (even if the Underwriters were treated as one entity for such purpose) or by any other method of allocation
which does not take account of the equitable considerations referred to above in this subsection (d). The amount paid or payable by an
indemnified party as a result of the losses, claims, damages or liabilities (or actions in respect thereof) referred to above in this
subsection (d) shall be deemed to include any legal or other expenses reasonably incurred by such indemnified party in connection
with investigating or defending any such action or claim. Notwithstanding the provisions of this subsection (d), no Underwriter shall
be required to contribute any amount in excess of the amount by which the total price at which the Shares underwritten by it and distributed
to the public were offered to the public exceeds the amount of any damages which such Underwriter has otherwise been required to pay by
reason of such untrue or alleged untrue statement or omission or alleged omission. No person guilty of fraudulent misrepresentation (within
the meaning of Section 11(f) of the Act) shall be entitled to contribution from any person who was not guilty of such fraudulent
misrepresentation. The Underwriters' obligations in this subsection (d) to contribute are several in proportion to their respective
underwriting obligations and not joint.
(e) The
obligations of the Company under this Section 9 shall be in addition to any liability which the Company may otherwise have and shall
extend, upon the same terms and conditions, to each employee, officer and director of each Underwriter and each person, if any, who controls
any Underwriter within the meaning of the Act and each broker-dealer or other affiliate of any Underwriter; and the obligations of the
Underwriters under this Section 9 shall be in addition to any liability which the respective Underwriters may otherwise have and
shall extend, upon the same terms and conditions, to each officer and director of the Company (including any person who, with his or her
consent, is named in the Registration Statement as about to become a director of the Company) and to each person, if any, who controls
the Company within the meaning of the Act.
10. (a)
If any Underwriter shall default in its obligation to purchase the Shares that it has agreed to purchase hereunder at a Time of Delivery,
you may in your discretion arrange for you or another party or other parties to purchase such Shares on the terms contained herein. If
within thirty-six hours after such default by any Underwriter you do not arrange for the purchase of such Shares, then the Company shall
be entitled to a further period of thirty-six hours within which to procure another party or other parties satisfactory to you to purchase
such Shares on such terms. In the event that, within the respective prescribed periods, you notify the Company that you have so arranged
for the purchase of such Shares, or the Company notifies you that it has so arranged for the purchase of such Shares, you or the Company
shall have the right to postpone such Time of Delivery for a period of not more than seven days, in order to effect whatever changes may
thereby be made necessary in the Registration Statement or the Prospectus, or in any other documents or arrangements, and the Company
agrees to file promptly any amendments or supplements to the Registration Statement or the Prospectus which in your opinion may thereby
be made necessary. The term “Underwriter” as used in this Agreement shall include any person substituted under this Section with
like effect as if such person had originally been a party to this Agreement with respect to such Shares.
(b) If,
after giving effect to any arrangements for the purchase of the Shares of a defaulting Underwriter or Underwriters by you and the Company
as provided in subsection (a) above, the aggregate number of such Shares which remains unpurchased does not exceed one-eleventh of
the aggregate number of all the Shares to be purchased at such Time of Delivery, then the Company shall have the right to require each
non-defaulting Underwriter to purchase the number of Shares which such Underwriter agreed to purchase hereunder at such Time of Delivery
and, in addition, to require each non-defaulting Underwriter to purchase its pro rata share (based on the number of Shares which such
Underwriter agreed to purchase hereunder) of the Shares of such defaulting Underwriter or Underwriters for which such arrangements have
not been made; but nothing herein shall relieve a defaulting Underwriter from liability for its default.
(c) If,
after giving effect to any arrangements for the purchase of the Shares of a defaulting Underwriter or Underwriters by you and the Company
as provided in subsection (a) above, the aggregate number of such Shares which remains unpurchased exceeds one-eleventh of the aggregate
number of all the Shares to be purchased at such Time of Delivery, or if the Company shall not exercise the right described in subsection
(b) above to require non-defaulting Underwriters to purchase Shares of a defaulting Underwriter or Underwriters, then this Agreement
(or, with respect to the Second Time of Delivery, the obligations of the Underwriters to purchase and of the Company to sell the Optional
Shares) shall thereupon terminate, without liability on the part of any non-defaulting Underwriter or the Company, except for the expenses
to be borne by the Company and the Underwriters as provided in Section 7 hereof and the indemnity and contribution agreements in
Section 9 hereof; but nothing herein shall relieve a defaulting Underwriter from liability for its default.
11. The
respective indemnities, rights of contribution, agreements, representations, warranties and other statements of the Company and the several
Underwriters, as set forth in this Agreement or made by or on behalf of them, respectively, pursuant to this Agreement, shall remain in
full force and effect, regardless of any investigation (or any statement as to the results thereof) made by or on behalf of any Underwriter
or any director, officer, employee, broker-dealer, affiliate or controlling person of any Underwriter, or the Company, or any officer
or director or controlling person of the Company, and shall survive delivery of and payment for the Shares.
12. If
this Agreement shall be terminated pursuant to Section 10 hereof, the Company shall not then be under any liability to any Underwriter
except as provided in Sections 7 and 9 hereof; but, if for any other reason, any Shares are not delivered by or on behalf of the
Company as provided herein, or the Underwriters decline to purchase the Shares for any reason permitted under this Agreement, the Company
will reimburse the Underwriters through you for all out-of-pocket expenses approved in writing by you, including fees and disbursements
of counsel, reasonably incurred by the Underwriters in making preparations for the purchase, sale and delivery of the Shares not so delivered,
but the Company shall then be under no further liability to any Underwriter except as provided in Sections 7 and 9 hereof.
13. In
all dealings hereunder, the Representatives shall act on behalf of each of the Underwriters, and the parties hereto shall be entitled
to act and rely upon any statement, request, notice or agreement on behalf of any Underwriter made or given by you jointly or by Goldman
Sachs & Co. LLC and Jefferies LLC on behalf of you as the Representatives.
All statements, requests,
notices and agreements hereunder shall be in writing, and (A) if to the Underwriters shall be delivered or sent by mail, telex or
facsimile transmission to the Representatives (i) in care of Goldman Sachs & Co. LLC, at 200 West Street, New York, New
York 10282-2198, Attention: Registration Department, and (ii) in care of Jefferies LLC, at 520 Madison Avenue, New York, New York
10022, Attention: General Counsel; (B) if to the Company shall be delivered or sent by mail, telex or facsimile transmission to the
address of the Company set forth on the cover of the Registration Statement, Attention: Chief Executive Officer, with a copy to Goodwin
Procter LLP, 100 Northern Avenue, Boston, Massachusetts 02210, Attention: Kingsley L. Taft and Sarah Ashfaq; provided, however, that any
notice to an Underwriter pursuant to Section 9(c) hereof shall be delivered or sent by e-mail, mail, telex or facsimile transmission
to such Underwriter at its address set forth in its Underwriters' Questionnaire, or telex constituting such Questionnaire, which address
will be supplied to the Company by you upon request; provided further, however, that notices under subsection 5(e) shall be in writing.
Any such statements, requests, notices or agreements shall take effect upon receipt thereof.
In accordance with the requirements
of the USA Patriot Act (Title III of Pub. L. 107-56 (signed into law October 26, 2001)), the Underwriters are required to obtain,
verify and record information that identifies their respective clients, including the Company, which information may include the name
and address of their respective clients, as well as other information that will allow the Underwriters to properly identify their respective
clients.
14. This
Agreement shall be binding upon, and inure solely to the benefit of, the Underwriters, the Company and, to the extent provided in Sections
9 and 11 hereof, the officers and directors of the Company and each person who controls the Company or any Underwriter, or any director,
officer, employee, broker-dealer or affiliate of any Underwriter, and their respective heirs, executors, administrators, successors and
assigns, and no other person shall acquire or have any right under or by virtue of this Agreement. No purchaser of any of the Shares from
any Underwriter shall be deemed a successor or assign by reason merely of such purchase.
15. Time
shall be of the essence of this Agreement. As used herein, the term “business day” shall mean any day when the Commission's
office in Washington, D.C. is open for business.
16. The
Company acknowledges and agrees that (i) the purchase and sale of the Shares pursuant to this Agreement is an arm’s-length
commercial transaction between the Company, on the one hand, and the several Underwriters, on the other, and does not constitute a recommendation,
investment advice, or solicitation of any action by the Underwriters, (ii) in connection therewith and with the process leading to
such transaction each Underwriter is acting solely as a principal and not the agent or fiduciary of the Company, (iii) no Underwriter
has assumed an advisory or fiduciary responsibility in favor of the Company with respect to the offering contemplated hereby or the process
leading thereto (irrespective of whether such Underwriter has advised or is currently advising the Company on other matters) or any other
obligation to the Company except the obligations expressly set forth in this Agreement, (iv) the Underwriters and their respective
affiliates may be engaged in a broad range of transactions that involve interests that differ from those of the Company, (iv) the
Underwriters have not provided any legal, accounting, regulatory, investment or tax advice with respect to the offering of the Shares
and the Company has consulted its own legal, accounting, financial, regulatory and tax advisors to the extent it deemed appropriate, and
(v) none of the activities of the Underwriters in connection with the transactions contemplated herein constitutes a recommendation,
investment advice, or solicitation of any action by the Underwriters with respect to any entity or natural person. The Company agrees
that it will not claim that the Underwriters, or any of them, has rendered advisory services of any nature or respect, or owes a fiduciary
or similar duty to the Company, in connection with such transaction or the process leading thereto.
17. This
Agreement supersedes all prior agreements and understandings (whether written or oral) between the Company and the Underwriters, or any
of them, with respect to the subject matter hereof.
18. This
Agreement and any transaction contemplated by this Agreement and any claim, controversy or dispute arising under or related thereto shall
be governed by and construed in accordance with the laws of the State of New York without regard to principles of conflict of laws that
would result in the application of any other law than the laws of the State of New York. The Company agrees that any suit or proceeding
arising in respect of this Agreement or any transaction contemplated by this Agreement will be tried exclusively in the U.S. District
Court for the Southern District of New York or, if that court does not have subject matter jurisdiction, in any state court located in
The City and County of New York and the Company agrees to submit to the jurisdiction of, and to venue in, such courts.
19. The
Company and each of the Underwriters hereby irrevocably waives, to the fullest extent permitted by applicable law, any and all right to
trial by jury in any legal proceeding arising out of or relating to this Agreement or the transactions contemplated hereby.
20. This
Agreement may be executed by any one or more of the parties hereto in any number of counterparts, each of which shall be deemed to be
an original, but all such counterparts shall together constitute one and the same instrument. Counterparts may be delivered via facsimile,
electronic mail (including any electronic signature covered by the U.S. federal ESIGN Act of 2000, Uniform Electronic Transactions Act,
the Electronic Signatures and Records Act or other applicable law, e.g., www.docusign.com) or other transmission method and any counterpart
so delivered shall be deemed to have been duly and validly delivered and be valid and effective for all purposes.
21. Notwithstanding anything
herein to the contrary, the Company is authorized to disclose to any persons the U.S. federal and state income tax treatment and tax structure
of the potential transaction and all materials of any kind (including tax opinions and other tax analyses) provided to the Company relating
to that treatment and structure, without the Underwriters imposing any limitation of any kind. However, any information relating to the
tax treatment and tax structure shall remain confidential (and the foregoing sentence shall not apply) to the extent necessary to enable
any person to comply with securities laws. For this purpose, “tax structure” is limited to any facts that may be relevant
to that treatment.
22. Recognition of the U.S.
Special Resolution Regimes.
(a) In
the event that any Underwriter that is a Covered Entity becomes subject to a proceeding under a U.S. Special Resolution Regime, the transfer
from such Underwriter of this Agreement, and any interest and obligation in or under this Agreement, will be effective to the same extent
as the transfer would be effective under the U.S. Special Resolution Regime if this Agreement, and any such interest and obligation, were
governed by the laws of the United States or a state of the United States.
(b) In the event that
any Underwriter that is a Covered Entity or a BHC Act Affiliate of such Underwriter becomes subject to a proceeding under a U.S. Special
Resolution Regime, Default Rights under this Agreement that may be exercised against such Underwriter are permitted to be exercised to
no greater extent than such Default Rights could be exercised under the U.S. Special Resolution Regime if this Agreement were governed
by the laws of the United States or a state of the United States.
(c) As used in this section:
“BHC Act Affiliate”
has the meaning assigned to the term “affiliate” in, and shall be interpreted in accordance with, 12 U.S.C. § 1841(k).
“Covered Entity”
means any of the following:
(i) a “covered
entity” as that term is defined in, and interpreted in accordance with, 12 C.F.R. § 252.82(b);
(ii) a “covered
bank” as that term is defined in, and interpreted in accordance with, 12 C.F.R. § 47.3(b); or
(iii) a “covered
FSI” as that term is defined in, and interpreted in accordance with, 12 C.F.R. § 382.2(b).
“Default Right”
has the meaning assigned to that term in, and shall be interpreted in accordance with, 12 C.F.R. §§ 252.81, 47.2 or 382.1, as
applicable.
“U.S. Special Resolution
Regime” means each of (i) the Federal Deposit Insurance Act and the regulations promulgated thereunder and (ii) Title
II of the Dodd-Frank Wall Street Reform and Consumer Protection Act and the regulations promulgated thereunder.
If the foregoing is in accordance
with your understanding, please sign and return to us one for the Company and each of the Representatives plus one for each counsel counterparts
hereof, and upon the acceptance hereof by you, on behalf of each of the Underwriters, this letter and such acceptance hereof shall constitute
a binding agreement between each of the Underwriters and the Company. It is understood that your acceptance of this letter on behalf of
each of the Underwriters is pursuant to the authority set forth in a form of Agreement among Underwriters, the form of which shall be
submitted to the Company for examination upon request, but without warranty on your part as to the authority of the signers thereof.
[Signature Page Follows]
| |
Very truly yours, |
| |
|
| |
City Therapeutics, Inc. |
| |
|
| |
By: |
|
| |
Name: |
|
| |
Title: |
|
| Accepted as of the date hereof: |
|
| |
|
| Goldman Sachs & Co. LLC |
|
| |
|
| By: |
|
|
| Name: |
|
|
| Title: |
|
|
| |
|
| Jefferies LLC |
|
| |
|
| By: |
|
|
| Name: |
|
|
| Title: |
|
|
| |
|
| On behalf of each of the Underwriters |
|
SCHEDULE I
| Underwriter | |
Total
Number of
Firm Shares
to be
Purchased | |
Number
of
Optional Shares
to be
Purchased if
Maximum
Option
Exercised |
| Goldman Sachs & Co. LLC | |
[·] | |
[·] |
| Jefferies LLC | |
[·] | |
[·] |
| Stifel, Nicolaus & Company, Incorporated | |
[·] | |
[·] |
| Oppenheimer & Co. Inc. | |
[·] | |
[·] |
| Total | |
[·] | |
[·] |
SCHEDULE II
(a) Issuer Free Writing Prospectuses not included in the Pricing
Disclosure Package:
Electronic Roadshow dated October 2026
(b) Additional Documents Incorporated by Reference:
[None]
(c) Information other than the Pricing Prospectus that comprise
the Pricing Disclosure Package:
The initial public offering price per share for
the Shares is $[·].
The number of Shares purchased by the Underwriters
is [·].
(d) Written
Testing-the-Waters Communications:
Testing-the-Waters investor deck dated July 2026
Testing-the-Waters investor deck dated August 2026
ANNEX I
FORM OF PRESS RELEASE
City Therapeutics, Inc.
[Date]
City Therapeutics, Inc. (the “Company”) announced
today that Goldman Sachs & Co. LLC and Jefferies LLC, the lead book-running managers in the Company’s recent public sale
of shares of common stock, are [waiving] [releasing] a lock-up restriction with respect to shares of the Company’s common stock
held by [certain officers or directors] [an officer or director] of the Company. The [waiver] [release] will take effect on [date], and
the shares may be sold on or after such date.
This press release is not an offer for sale of the securities in
the United States or in any other jurisdiction where such offer is prohibited, and such securities may not be offered or sold in the United
States absent registration or an exemption from registration under the United States Securities Act of 1933, as amended.
ANNEX II
FORM OF LOCK-UP AGREEMENT
City Therapeutics, Inc.
Lock-Up Agreement
,
2026
Goldman Sachs & Co. LLC
Jefferies LLC
As Representatives of the several Underwriters
named in Schedule I to the Underwriting Agreement
c/o Goldman Sachs & Co. LLC
200 West Street
New York, NY 10282-2198
c/o Jefferies LLC
520 Madison Avenue
New York, NY 10022
Re: City Therapeutics, Inc. - Lock-Up Agreement
Ladies and Gentlemen:
The undersigned understands
that you, as representatives (the “Representatives”), propose to enter into an underwriting agreement (the “Underwriting
Agreement”) on behalf of the several Underwriters named in Schedule I to such agreement (collectively, the “Underwriters”),
with City Therapeutics, Inc., a Delaware corporation (the “Company”), providing for a public offering (the “Public
Offering”) of shares (the “Shares”) of common stock, par value $0.0001 per share, of the Company (the “Common
Stock”) pursuant to a Registration Statement on Form S-1 (the “Registration Statement”) to be filed with the Securities
and Exchange Commission (the “SEC”).
In consideration of the agreement
by the Underwriters to offer and sell the Shares, and of other good and valuable consideration the receipt and sufficiency of which is
hereby acknowledged, the undersigned agrees that, during the period beginning from the date of this Lock-Up Agreement and continuing to
and including the date 180 days after the date of the final prospectus relating to the Public Offering (the “Prospectus”)
(such period, the “Lock-Up Period”), the undersigned shall not, and shall not cause or direct any of its affiliates to, (i) offer,
sell, contract to sell, pledge, grant any option, right or warrant to purchase, purchase any option or contract to sell, lend or otherwise
transfer or dispose of any shares of Common Stock, or any options or warrants to purchase any shares of Common Stock, or any securities
convertible into, exchangeable for or that represent the right to receive shares of Common Stock (such shares of Common Stock, options,
rights, warrants or other securities, collectively, “Lock-Up Securities”), including without limitation any such Lock-Up Securities
now owned or hereafter acquired by the undersigned, (ii) engage in any hedging or other transaction or arrangement (including, without
limitation, any short sale or the purchase or sale of, or entry into, any put or call option, or combination thereof, forward, swap or
any other derivative transaction or instrument, however described or defined) which is designed to or which reasonably could be expected
to lead to or result in a sale, loan, pledge or other disposition (whether by the undersigned or someone other than the undersigned),
or transfer of any of the economic consequences of ownership, in whole or in part, directly or indirectly, of any Lock-Up Securities,
whether any such transaction or arrangement (or instrument provided for thereunder) would be settled by delivery of Common Stock or other
securities, in cash or otherwise (any such sale, loan, pledge or other disposition, or transfer of economic consequences, a “Transfer”),
(iii) make any demand for or exercise any right with respect to the registration of any Lock-Up Securities or (iv) otherwise
publicly announce any intention to engage in or cause any action, activity, transaction or arrangement described in clause (i), (ii) or
(iii) above. The undersigned represents and warrants that the undersigned is not, and has not caused or directed any of its affiliates
to be or become, currently a party to any agreement or arrangement that provides for, is designed to or reasonably could be expected to
lead to or result in any Transfer during the Lock-Up Period.
Notwithstanding the foregoing,
the undersigned may:
| (a) | transfer the undersigned’s Lock-Up Securities; |
| i. | as one or more bona fide gifts or charitable contributions, or for bona fide estate planning
purposes; |
| ii. | upon death by will, testamentary document or intestate succession; |
| iii. | if the undersigned is a natural person, to any member of the undersigned’s immediate family (for
purposes of this Lock-Up Agreement, “immediate family” shall mean any relationship by blood, current or former marriage, domestic
partnership, civil union or adoption, not more remote than first cousin) or to any trust for the direct or indirect benefit of the undersigned
or the immediate family of the undersigned or, if the undersigned is a trust, to a trustor, trustee (or co-trustee) or beneficiary of
the trust or the estate of a beneficiary of such trust; |
| iv. | to a corporation, partnership, limited liability company, investment fund or other entity (A) of
which the undersigned and the immediate family of the undersigned are the legal and beneficial owner of all of the outstanding equity
securities or similar interests or (B) controlled by, or under common control with, the undersigned or the immediate family of the
undersigned; |
| v. | to a nominee or custodian of a person or entity to whom a disposition or transfer would be permissible
under clauses (a)(i) through (iv) above; |
| vi. | if the undersigned is a corporation, partnership, limited liability company, trust or other business entity,
(A) to another corporation, partnership, limited liability company, trust or other business entity that is a subsidiary or an affiliate
(as defined in Rule 405 under the Securities Act of 1933, as amended) of the undersigned, or to any investment fund or other entity
which fund or entity is controlled or managed by or under common control or common investment management with the undersigned or affiliates
of the undersigned (including, for the avoidance of doubt, where the undersigned is a partnership, to its general partner or a successor
partnership or fund, or any other funds managed by such partnership), or (B) as part of a disposition, transfer or distribution by
the undersigned to its stockholders, partners, members or other equityholders or to the estate of any such stockholders, partners, members
or other equityholders; |
| vii. | by operation of law, such as pursuant to a qualified domestic order, divorce settlement, divorce decree
or separation agreement; |
| viii. | to the Company upon death or disability of the undersigned, or, if the undersigned is an employee of the
Company or subsidiaries, upon death, disability or termination of employment, in each case, of such employee; |
| ix. | if the undersigned is not an officer or director of the Company, in connection with a sale of the undersigned’s
shares of Common Stock acquired (A) from the Underwriters in the Public Offering or (B) in open market transactions after the
closing date of the Public Offering; |
| x. | to the Company in connection with the vesting, settlement or exercise of restricted stock units, options,
warrants or other rights or instruments to purchase or acquire shares of Common Stock (including, in each case, by way of “net”
or “cashless” exercise) including any transfer to the Company for the payment of tax withholdings or remittance payments due
as a result of the vesting, settlement or exercise of such restricted stock units, options, warrants or other rights or instruments, or
in connection with the conversion of convertible securities, in all such cases pursuant to equity awards granted under a stock incentive
plan or other equity award plan, or pursuant to the terms of convertible securities, each as described in the Registration Statement,
the preliminary prospectus relating to the Shares included in the Registration Statement immediately prior to the time the Underwriting
Agreement is executed and the Prospectus, provided that any securities received upon such vesting, settlement, exercise or conversion
shall be subject to the terms of this Lock-Up Agreement; or |
| xi. | with the prior written consent of Goldman Sachs & Co. LLC and Jefferies LLC on behalf of the
Underwriters; |
provided that (A) in the case
of clauses (a)(i), (ii), (iii), (iv), (v) and (vi) above, such transfer or distribution shall not involve a disposition for
value, (B) in the case of clauses (a)(i), (ii), (iii), (iv), (v), (vi) and (vii) above, it shall be a condition to the
transfer or distribution that the donee, devisee, transferee or distributee, as the case may be, shall sign and deliver a lock-up agreement
in the form of this Lock-Up Agreement, (C) in the case of clauses (a)(ii), (iii), (iv), (v) and (vi) above, no filing by
any party (including, without limitation, any donor, donee, devisee, transferor, transferee, distributor or distributee) under the Securities
Exchange Act of 1934, as amended (the “Exchange Act”), or other public filing, report or announcement reporting a reduction
in beneficial ownership of Lock-Up Securities shall be required or shall be voluntarily made in connection with such transfer or distribution
(other than a filing on a Form 5 or a filing required pursuant to Section 13 of the Exchange Act and the rules and regulations
promulgated thereunder, in each case, made after the expiration of the Lock-Up Period referred to above), and (D) in the case of
clauses (a)(i), (vii), (viii), (ix) and (x) above, no filing under the Exchange Act or other public filing, report or announcement
shall be voluntarily made, and if any such filing, report or announcement shall be legally required during the Lock-Up Period, such filing,
report or announcement shall clearly indicate in the footnotes thereto (A) the circumstances of such transfer or distribution and
(B) in the case of a transfer or distribution pursuant to clauses (a)(i) or (vii) above, that the donee, devisee, transferee
or distributee has agreed to be bound by a lock-up agreement in the form of this Lock-Up Agreement;
| (b) | transfer the undersigned’s Lock-Up Securities to the Company pursuant to an agreement under which
the Company has the option to repurchase shares or a right of first refusal with respect to transfer of such shares, provided that no
filing under the Exchange Act or other public filing, report or announcement shall be voluntarily made, and if any such filing, report
or announcement shall be legally required during the Lock-Up Period, such filing, report or announcement shall clearly indicate in the
footnotes thereto the circumstances of such transfer or distribution; |
| (c) | enter into a written plan meeting the requirements of Rule 10b5-1 under the Exchange Act relating
to the transfer, sale or other disposition of the undersigned’s Lock-Up Securities, if then permitted by the Company, provided that
none of the securities subject to such plan may be transferred, sold or otherwise disposed of until after the expiration of the Lock-Up
Period and no public announcement, report or filing under the Exchange Act, or any other public filing, report or announcement, shall
be voluntarily made regarding the establishment of such plan during the Lock-Up Period, and if any such filing, report or announcement
shall be legally required during the Lock-Up Period, such filing, report or announcement shall clearly indicate in the footnotes thereto
that that none of the securities subject to such plan may be transferred, sold or otherwise disposed of pursuant to such plan until after
the expiration of the Lock-Up Period; |
| (d) | transfer the undersigned’s Lock-Up Securities pursuant to a bona fide third-party tender offer,
merger, consolidation or other similar transaction that is approved by the Board of Directors of the Company and made to all holders of
the Company’s capital stock involving a Change of Control of the Company (for purposes hereof, “Change of Control” shall
mean the transfer (whether by tender offer, merger, consolidation or other similar transaction), in one transaction or a series of related
transactions, to a person or group of affiliated persons, of shares of capital stock if, after such transfer, such person or group of
affiliated persons would hold at least a majority of the outstanding voting securities of the Company (or the surviving entity)) and (ii) enter
into any lock-up, voting or similar agreement pursuant to which the undersigned may agree to transfer, sell, tender or otherwise dispose
of Common Stock or such other securities in connection with a transaction described in clause (i) above; provided that in the event
that such tender offer, merger, consolidation or other similar transaction is not completed, the undersigned’s Lock-Up Securities
shall remain subject to the provisions of this Lock-Up Agreement; and |
| (e) | convert outstanding preferred stock of the Company, warrants to acquire preferred stock or convertible
securities into shares of Common Stock or warrants to acquire shares of Common Stock, provided that any Lock-Up Securities received upon
such conversion shall remain subject to the provisions of this Lock-Up Agreement. |
If the undersigned is an officer
or director of the Company, the undersigned further agrees that the foregoing provisions shall be equally applicable to any issuer-directed
or other Shares the undersigned may purchase in the Public Offering.
If the undersigned is not
a natural person, the undersigned represents and warrants that no single natural person, entity or “group” (within the meaning
of Section 13(d)(3) of the Exchange Act), other than a natural person, entity or “group” (as described above) that
has executed a Lock-Up Agreement in substantially the same form as this Lock-Up Agreement, beneficially owns, directly or indirectly,
50% or more of the common equity interests, or 50% or more of the voting power, in the undersigned.
If the undersigned is an officer
or director of the Company, (i) Goldman Sachs & Co. LLC and Jefferies LLC agree that, at least three business days before
the effective date of any release or waiver of the foregoing restrictions in connection with a transfer of Lock-Up Securities, Goldman
Sachs & Co. LLC and Jefferies LLC will notify the Company of the impending release or waiver, and (ii) the Company has agreed
in the Underwriting Agreement to announce the impending release or waiver by press release through a major news service (or such other
method approved by Goldman Sachs & Co. LLC and Jefferies LLC that satisfies the requirements of FINRA Rule 5131(d)(2)) at
least two business days before the effective date of the release or waiver. Any release or waiver granted by Goldman Sachs &
Co. LLC and Jefferies LLC hereunder to any such officer or director shall only be effective two business days after the publication date
of such announcement. The provisions of this paragraph will not apply if (i) the release or waiver is effected solely to permit a
transfer not for consideration or that is to an immediate family member as defined in FINRA Rule 5130(i)(5) and (ii) the
transferee has agreed in writing to be bound by the same terms described in this Lock-Up Agreement to the extent and for the duration
that such terms remain in effect at the time of the transfer.
The undersigned now has, and,
except as contemplated by clauses (a) and (d) of the third paragraph of this Lock-Up Agreement, for the duration of this Lock-Up
Agreement will have, good and marketable title to the undersigned’s Lock-Up Securities, free and clear of all liens, encumbrances
and claims whatsoever. The undersigned also agrees and consents to the entry of stop transfer instructions with the Company’s transfer
agent and registrar against the transfer of the undersigned’s Lock-Up Securities except in compliance with the foregoing restrictions.
The undersigned acknowledges
and agrees that none of the Underwriters has made any recommendation or provided any investment or other advice to the undersigned with
respect to this Lock-Up Agreement or the subject matter hereof, and the undersigned has consulted its own legal, accounting, financial,
regulatory, tax and other advisors with respect to this Lock-Up Agreement and the subject matter hereof to the extent the undersigned
has deemed appropriate. The undersigned further acknowledges and agrees that, although the Underwriters may have provided or hereafter
provide to the undersigned in connection with the Public Offering a Form CRS and/or certain other disclosures as contemplated by
Regulation Best Interest, the Underwriters have not made and are not making a recommendation to the undersigned to enter into this Lock-Up
Agreement or to transfer, sell or dispose of, or to refrain from transferring, selling or disposing of, any shares of Common Stock, and
nothing set forth in such disclosures or herein is intended to suggest that any Underwriter is making such a recommendation.
This Lock-Up Agreement shall
automatically terminate and the undersigned shall be released from all of his, her or its obligations hereunder upon the earlier of (i) the
date on which the Registration Statement filed with the SEC with respect to the Public Offering is withdrawn, (ii) the date on which
for any reason the Underwriting Agreement is terminated (other than the provisions thereof that survive termination) prior to payment
for and delivery of the Shares to be sold thereunder (other than pursuant to the Underwriters’ option thereunder to purchase additional
Shares), (iii) the date on which the Company notifies the Representatives, in writing and prior to the execution of the Underwriting
Agreement, that it does not intend to proceed with the Public Offering and (iv) January 31, 2027, in the event that the Underwriting
Agreement has not been executed by such date (provided, however, that the Company may, by written notice to the undersigned prior to such
date, extend such date by a period of up to an additional 90 days).
The undersigned understands
that the Company and the Underwriters are relying upon this Lock-Up Agreement in proceeding toward consummation of the Public Offering.
The undersigned further understands that this Lock-Up Agreement is irrevocable and shall be binding upon the undersigned’s heirs,
legal representatives, successors and assigns. The undersigned hereby represents and warrants that the undersigned has full power and
authority to enter into this Lock-Up Agreement. This Lock-Up Agreement shall be governed by, and construed in accordance with, the laws
of the State of New York, without regard to principles of conflict of laws that would result in the application of any law other than
the laws of the State of New York. This Lock-Up Agreement may be delivered via facsimile, electronic mail (including pdf or any electronic
signature complying with the U.S. federal ESIGN Act of 2000, e.g., www.docusign.com or www.echosign.com) or other transmission method,
and any counterpart so delivered shall be deemed to have been duly and validly delivered and be valid and effective for all purposes.
[Signature Page Follows]
Very truly yours,
| IF AN INDIVIDUAL: |
|
IF AN ENTITY: |
| |
|
|
|
| By: |
|
|
|
| |
|
(duly authorized signature) |
|
(please print complete name of entity) |
| |
|
|
| Name: |
|
|
By: |
|
| |
|
(please print full name) |
|
|
|
(duly authorized signature) |
| |
|
|
| |
|
Name: |
|
| |
|
|
|
(please print full name) |
| |
|
|
| |
|
Title: |
|
| |
|
|
|
(please print full title) |
Exhibit 3.1
AMENDED AND RESTATED
CERTIFICATE OF INCORPORATION
OF
CITY THERAPEUTICS, INC.
(Pursuant to Sections 242 and 245 of the
General Corporation Law of the State of Delaware)
City Therapeutics, Inc., a
corporation organized and existing under and by virtue of the provisions of the General Corporation Law of the State of Delaware (the
“General Corporation Law”),
DOES HEREBY CERTIFY:
1.
That the name of this corporation is City Therapeutics, Inc., and that this corporation was originally incorporated pursuant
to the General Corporation Law on July 17, 2023.
2.
That the Board of Directors of this corporation (the “Board of Directors”) duly adopted resolutions proposing
to amend and restate the Certificate of Incorporation of this corporation, declaring said amendment and restatement to be advisable and
in the best interests of this corporation and its stockholders, and authorizing the appropriate officers of this corporation to solicit
the consent of the stockholders therefor, which resolution setting forth the proposed amendment and restatement is as follows:
RESOLVED,
that the Certificate of Incorporation of this corporation be amended and restated in its entirety to read as follows:
First:
The name of this corporation is City Therapeutics, Inc. (the “Corporation”).
Second:
The address of the registered office of the Corporation in the State of Delaware is 3500 South DuPont Highway in the City of Dover, County
of Kent, 19901. The name of its registered agent at such address is Incorporating Services, Ltd.
Third:
The nature of the business or purposes to be conducted or promoted is to engage in any lawful act or activity for which corporations may
be organized under the General Corporation Law.
Fourth:
The total number of shares of all classes of stock which the Corporation shall have authority to issue is (i) 41,000,000 shares of Common
Stock, $0.0001 par value per share (“Common Stock”) and (ii) 26,266,656 shares of Preferred Stock, $0.0001 par value
per share (“Preferred Stock”).
The following is a statement
of the designations and the powers, privileges and rights, and the qualifications, limitations or restrictions thereof in respect of each
class of capital stock of the Corporation.
A.
COMMON STOCK
1.
General. The voting, dividend and liquidation rights of the holders of the Common Stock are subject to and qualified by
the rights, powers and preferences of the holders of the Preferred Stock set forth herein.
2.
Voting. The holders of the Common Stock are entitled to one vote for each share of Common Stock held at all meetings of
stockholders (and written actions in lieu of meetings); provided, however, that, except as otherwise required by law, holders
of Common Stock, as such, shall not be entitled to vote on any amendment to this Amended and Restated Certificate of Incorporation that
relates solely to the terms of one or more outstanding series of Preferred Stock if the holders of such affected series are entitled,
either separately or together with the holders of one or more other such series, to vote thereon pursuant to this Amended and Restated
Certificate of Incorporation or pursuant to the General Corporation Law. There shall be no cumulative voting. The number of authorized
shares of Common Stock may be increased or decreased (but not below the number of shares thereof then outstanding) by (in addition to
any vote of the holders of one or more series of Preferred Stock that may be required by the terms of this Amended and Restated Certificate
of Incorporation) the affirmative vote of the holders of shares of capital stock of the Corporation representing a majority of the votes
represented by all outstanding shares of capital stock of the Corporation entitled to vote, irrespective of the provisions of Section
242(b)(2) of the General Corporation Law.
B.
PREFERRED STOCK
4,544,990
shares of the authorized Preferred Stock of the Corporation are hereby designated “Series Seed Preferred Stock”, 13,996,318
shares of the authorized Preferred Stock of the Corporation are hereby designated “Series A Preferred Stock”, 924,030
shares of the authorized Preferred Stock of the Corporation are hereby designated “Series B-1 Preferred Stock” and
6,801,318 shares of the authorized Preferred Stock of the Corporation are hereby designated “Series B Preferred Stock”,
with the following rights, preferences, powers, privileges and restrictions, qualifications and limitations. Unless otherwise indicated,
references to “Sections” in this Part B of this Article Fourth refer to sections of Part B of this Article Fourth.
References to “Preferred Stock” mean, collectively, the Series Seed Preferred Stock, Series A Preferred Stock, Series
B-1 Preferred Stock and Series B Preferred Stock.
1.
Dividends. The holders of then outstanding shares of Preferred Stock shall be entitled to receive, only when, as and if
declared by the Board of Directors, out of any funds and assets legally available therefor, dividends at the rate of 8% of the Original
Issue Price (as defined below) for each share of Preferred Stock, prior and in preference to any declaration or payment of any other dividend
(other than dividends on shares of Common Stock payable in shares of Common Stock). The right to receive dividends on shares of Preferred
Stock pursuant to the preceding sentence of this Section 1 shall not be cumulative, and no right to dividends shall accrue
to holders of Preferred Stock by reason of the fact that dividends on said shares are not declared. The Corporation shall not declare,
pay or set aside any dividends on shares of any other class or series of capital stock of the Corporation (other than dividends on shares
of Common Stock payable in shares of Common Stock) unless (in addition to the obtaining of any consents required elsewhere in this Amended
and Restated Certificate of Incorporation) the holders of the Preferred Stock then outstanding shall first receive, or simultaneously
receive, in addition to the dividends payable pursuant to the first sentence of this Section 1, a dividend on each outstanding
share of Preferred Stock in an amount at least equal to (i) in the case of a dividend on Common Stock or any class or series that is convertible
into Common Stock, that dividend per share of Preferred Stock as would equal the product of (A) the dividend payable on each share of
such class or series determined, if applicable, as if all shares of such class or series had been converted into Common Stock and (B)
the number of shares of Common Stock issuable upon conversion of a share of Preferred Stock, in each case calculated on the record date
for determination of holders entitled to receive such dividend or (ii) in the case of a dividend on any class or series that is not convertible
into Common Stock, at a rate per share of Preferred Stock determined by (A) dividing the amount of the dividend payable on each share
of such class or series of capital stock by the original issuance price of such class or series of capital stock (subject to appropriate
adjustment in the event of any stock dividend, stock split, combination or other similar recapitalization with respect to such class or
series) and (B) multiplying such fraction by an amount equal to the Original Issue Price (as defined below); provided that, if
the Corporation declares, pays or sets aside, on the same date, a dividend on shares of more than one class or series of capital stock
of the Corporation, the dividend payable to the holders of Preferred Stock pursuant to this Section 1 shall be calculated based
upon the dividend on the class or series of capital stock that would result in the highest Preferred Stock dividend. The “Original
Issue Price” shall mean, with respect to the Series Seed Preferred Stock, $1.0000 per share, with respect to the Series A Preferred
Stock, $10.0295 per share, with respect to the Series B-1 Preferred Stock, $12.4351 per share and with respect to the Series B Preferred
Stock, $14.6295 per share, each subject to appropriate adjustment in the event of any stock dividend, stock split, combination or other
similar recapitalization with respect to the applicable Preferred Stock.
2.
Liquidation, Dissolution or Winding Up; Certain Mergers, Consolidations and Asset Sales.
2.1
Preferential Payments to Holders of Preferred Stock. In the event of any voluntary or involuntary liquidation, dissolution
or winding up of the Corporation, the holders of shares of Preferred Stock then outstanding shall be entitled to be paid out of the assets
of the Corporation available for distribution to its stockholders, and in the event of a Deemed Liquidation Event (as defined below),
the holders of shares of Preferred Stock then outstanding shall be entitled to be paid out of the consideration payable to stockholders
in such Deemed Liquidation Event or out of the Available Proceeds (as defined below), as applicable, on a pari passu basis based
on their relative Liquidation Amounts (as defined below) and before any payment shall be made to the holders of Common Stock by reason
of their ownership thereof, an amount per share of each such series of Preferred Stock equal to the applicable Original Issue Price for
such series of Preferred Stock, plus any dividends declared but unpaid thereon (the amount payable pursuant to this sentence is hereinafter
referred to as the “Liquidation Amount”). If upon any such liquidation, dissolution or winding up of the Corporation
or Deemed Liquidation Event, the assets of the Corporation available for distribution to its stockholders shall be insufficient to pay
the holders of shares of Preferred Stock the full amount to which they shall be entitled under this Section 2.1, the holders
of shares of Preferred Stock shall share ratably in any distribution of the assets available for distribution in proportion to the respective
amounts which would otherwise be payable in respect of the shares held by them upon such distribution if all amounts payable on or with
respect to such shares were paid in full.
2.2
Payments to Holders of Common Stock. In the event of any voluntary or involuntary liquidation, dissolution or winding up
of the Corporation, after the payment in full of all Liquidation Amounts required to be paid to the holders of shares of Preferred Stock,
the remaining assets of the Corporation available for distribution to its stockholders or, in the case of a Deemed Liquidation Event,
the consideration not payable to the holders of shares of Preferred Stock pursuant to Section 2.1 or the remaining Available Proceeds,
as the case may be, shall be distributed among the holders of shares of Common Stock, pro rata based on the number of shares of Common
Stock held by each such holder. Notwithstanding the above, for purposes of determining the amount each holder of shares of Preferred Stock
is entitled to receive with respect to a Deemed Liquidation Event, liquidation, dissolution or winding up, each such holder of shares
of a series of Preferred Stock shall be deemed to have converted (regardless of whether such holder actually converted) such holder’s
shares of such series into shares of Common Stock immediately prior to the Deemed Liquidation Event, liquidation, dissolution or winding
up if, as a result of an actual conversion, such holder would receive, in the aggregate, an amount greater than the amount that would
be distributed to such holder if such holder did not convert such series of Preferred Stock into shares of Common Stock. If any such holder
shall be deemed to have converted shares of Preferred Stock into Common Stock pursuant to this paragraph, then such holder shall not be
entitled to receive any distribution that would otherwise be made to holders of Preferred Stock that have not converted (or have not been
deemed to have converted) into shares of Common Stock.
2.3
Deemed Liquidation Events.
2.3.1 Definition. Each of the following events shall be considered a “Deemed Liquidation Event” unless the
holders of at least a majority of the outstanding shares of Preferred Stock, voting together as a single class on an as-converted to Common
Stock basis, which majority must include the Specified Holders (as defined in that certain Amended and Restated Voting Agreement, dated
as of May 20, 2026, by and among the Corporation and the other parties thereto, as may be amended from time to time (the “Voting
Agreement”)) (the “Requisite Holders”) elect otherwise by written notice sent to the Corporation at least
five days prior to the effective date of any such event:
(a) a
merger, consolidation, statutory conversion, transfer of the Corporation, domestication, or continuance in which
| (i) | the Corporation is a constituent party or |
| (ii) | a subsidiary of the Corporation is a constituent party and the Corporation issues shares of its capital
stock pursuant to such merger, consolidation, statutory conversion, transfer of the Corporation, domestication, or continuance, |
except any such merger, consolidation, statutory
conversion, transfer of the Corporation, domestication, or continuance involving the Corporation or a subsidiary in which the shares of
capital stock of the Corporation outstanding immediately prior to such merger, consolidation, statutory conversion, transfer of the Corporation,
domestication, or continuance continue to represent, or are converted into or exchanged for shares of capital stock or other equity interests
that represent, immediately following such merger, consolidation, statutory conversion, transfer of the Corporation, domestication, or
continuance, a majority, by voting power, of the capital stock or other equity interests of (1) the surviving or resulting corporation
or entity; or (2) if the surviving or resulting corporation or entity is a wholly owned subsidiary of another corporation or entity immediately
following such merger, consolidation, statutory conversion, transfer of the Corporation, domestication, or continuance, the parent corporation
or entity of such surviving or resulting corporation or entity; or
(b)
(1) the sale, lease, transfer, exclusive license or other disposition, in a single transaction or series of related transactions,
by the Corporation or any subsidiary of the Corporation of all or substantially all the assets of the Corporation and its subsidiaries
taken as a whole, or (2) the sale, lease, transfer, exclusive license or other disposition (whether by merger, consolidation, statutory
conversion, transfer of the Corporation, domestication, or continuance or otherwise, and whether in a single transaction or a series of
related transactions) of one or more subsidiaries of the Corporation if substantially all of the assets of the Corporation and its subsidiaries
taken as a whole are held by such subsidiary or subsidiaries, except where such sale, lease, transfer, exclusive license or other disposition
is to a wholly owned subsidiary of the Corporation.
Notwithstanding the foregoing,
the sale of shares of Preferred Stock or Common Stock in a bona fide financing transaction shall not be deemed a Deemed Liquidation Event.
2.3.2
Effecting a Deemed Liquidation Event.
(a)
The Corporation shall not have the power to effect a Deemed Liquidation Event referred to in Section 2.3.1(a)(i) unless
the agreement or plan with respect to such transaction, or terms of such transaction (any such agreement, plan or terms, the “Merger
Agreement”) provides that the consideration payable to the stockholders of the Corporation in such Deemed Liquidation Event
shall be allocated to the holders of capital stock of the Corporation in accordance with Sections 2.1 and 2.2.
(b)
In the event of a Deemed Liquidation Event referred to in Section 2.3.1(a)(ii) or 2.3.1(b), if the Corporation does
not effect a dissolution of the Corporation under the General Corporation Law within 90 days after such Deemed Liquidation Event, then
(i) the Corporation shall send a written notice to each holder of Preferred Stock no later than the 90th day after the Deemed
Liquidation Event advising such holders of their right (and the requirements to be met to secure such right) pursuant to the terms of
the following clause (ii) to require the redemption of such shares of Preferred Stock, and (ii) if the Requisite Holders so request in
a written instrument delivered to the Corporation not later than 120 days after such Deemed Liquidation Event, the Corporation shall use
the consideration received by the Corporation for such Deemed Liquidation Event (net of any retained liabilities associated with the assets
sold or technology licensed, as determined in good faith by the Board of Directors), together with any other assets of the Corporation
available for distribution to its stockholders, all to the extent permitted by Delaware law governing distributions to stockholders (the
“Available Proceeds”), on the 150th day after such Deemed Liquidation Event (such date being the “Redemption
Date”), to redeem all outstanding shares of Preferred Stock at a price per share equal to the applicable Liquidation Amount.
Notwithstanding the foregoing, in the event of a redemption pursuant to the preceding sentence, if the Available Proceeds are not sufficient
to redeem all outstanding shares of Preferred Stock, the Corporation shall redeem a pro rata portion of each holder’s shares of
Preferred Stock to the fullest extent of such Available Proceeds, based on the respective amounts which would otherwise be payable in
respect of the shares to be redeemed if the Available Proceeds were sufficient to redeem all such shares, and shall redeem the remaining
shares as soon as it may lawfully do so under Delaware law governing distributions to stockholders. On or before the applicable Redemption
Date, each holder of shares of Preferred Stock to be redeemed on such Redemption Date, unless such holder has previously exercised his,
her or its right to convert such shares as provided in Section 4, shall, if such holder’s shares are certificated, surrender
the certificate or certificates representing such shares (or, if such registered holder alleges that such certificate has been lost, stolen
or destroyed, a lost certificate affidavit and agreement reasonably acceptable to the Corporation to indemnify the Corporation against
any claim that may be made against the Corporation on account of the alleged loss, theft or destruction of such certificate (the “Lost
Certificate Documentation”)) to the Corporation, as requested in writing by the Corporation, and thereupon the applicable Liquidation
Amount for such shares shall be payable to the order of the person whose name appears on such certificate or certificates as the owner
thereof. In the event less than all of the shares of Preferred Stock represented by a certificate are redeemed, a new certificate representing
the unredeemed shares of Preferred Stock shall promptly be issued to such holder. Prior to the distribution or redemption provided for
in this Section 2.3.2(b), the Corporation shall not expend or dissipate the consideration received for such Deemed Liquidation
Event, except to discharge expenses incurred in connection with such Deemed Liquidation Event or in the ordinary course of business.
2.3.3
Amount Deemed Paid or Distributed. The amount deemed paid or distributed to the holders of capital stock of the Corporation
upon any such merger, consolidation, sale, transfer, exclusive license, other disposition or redemption shall be the cash or the value
of the property, rights or securities to be paid or distributed to such holders pursuant to such Deemed Liquidation Event. The value of
such property, rights or securities shall be determined in good faith by the Board of Directors, including a majority of the Preferred
Directors (as defined below) then seated.
2.3.4
Allocation of Escrow and Contingent Consideration. In the event of a Deemed Liquidation Event pursuant to Section 2.3.1(a)(i),
if any portion of the consideration payable to the stockholders of the Corporation is payable only upon satisfaction of contingencies
(the “Additional Consideration”), the Merger Agreement shall provide that (a) the portion of such consideration that
is not Additional Consideration (such portion, the “Initial Consideration”) shall be allocated among the holders of
capital stock of the Corporation in accordance with Sections 2.1 and 2.2 as if the Initial Consideration were the only consideration
payable in connection with such Deemed Liquidation Event; and (b) any Additional Consideration which becomes payable to the stockholders
of the Corporation upon satisfaction of such contingencies shall be allocated among the holders of capital stock of the Corporation in
accordance with Sections 2.1 and 2.2 after taking into account the previous payment of the Initial Consideration as part
of the same transaction. For the purposes of this Section 2.3.4, consideration placed into escrow or retained as a holdback to
be available for satisfaction of indemnification or similar obligations in connection with such Deemed Liquidation Event shall be deemed
to be Additional Consideration.
3.
Voting.
3.1
General. On any matter presented to the stockholders of the Corporation for their action or consideration at any meeting
of stockholders of the Corporation (or by written consent of stockholders in lieu of meeting), each holder of outstanding shares of Preferred
Stock shall be entitled to cast the number of votes equal to the number of whole shares of Common Stock into which the shares of Preferred
Stock held by such holder are convertible as of the record date for determining stockholders entitled to vote on such matter. Except as
provided by law or by the other provisions of this Amended and Restated Certificate of Incorporation, holders of Preferred Stock shall
vote together with the holders of Common Stock as a single class and on an as-converted to Common Stock basis.
3.2
Election of Directors. The holders of record of the shares of Preferred Stock, voting together as a class on an as-converted
basis, shall be entitled to elect three directors of the Corporation (each, a “Preferred Director”), and the holders
of record of the shares of Common Stock, voting together as a class, shall be entitled to elect two directors of the Corporation. Any
director elected as provided in the preceding sentence may be removed without cause by, and only by, the affirmative vote of the holders
of the shares of the class or series of capital stock entitled to elect such director or directors, given either at a special meeting
of such stockholders duly called for that purpose or pursuant to a written consent of stockholders. If the holders of shares of Common
Stock and Preferred Stock, as the case may be, fail to elect a sufficient number of directors to fill all directorships for which they
are entitled to elect directors, voting exclusively and as a separate class, pursuant to the first sentence of this Section 3.2,
then any directorship not so filled shall remain vacant until such time as such holders elect a person to fill such directorship by vote
or written consent in lieu of a meeting; and no such directorship may be filled by stockholders of the Corporation other than by the stockholders
of the Corporation that are entitled to elect a person to fill such directorship, voting exclusively and as a separate class. The holders
of record of the shares of Common Stock and of any other class or series of voting stock (including the Preferred Stock), exclusively
and voting together as a single class on an as-converted basis, shall be entitled to elect the balance of the total number of directors
of the Corporation. At any meeting held for the purpose of electing a director, the presence in person or by proxy of the holders of a
majority of the outstanding shares of the class or series entitled to elect such director shall constitute a quorum for the purpose of
electing such director. Except as otherwise provided in this Section 3.2, a vacancy in any directorship filled by the holders
of any class or classes or series shall be filled only by vote or written consent in lieu of a meeting of the holders of such class or
classes or series or by any remaining director or directors elected by the holders of such class or classes or series pursuant to this
Section 3.2. Notwithstanding the provisions of Section 223(a)(1) and 223(a)(2) of the General Corporation Law, any vacancy,
including newly created directorships resulting from any increase in the authorized number of directors or amendment of the Certificate
of Incorporation, and vacancies created by removal or resignation of a director, may be filled by a majority of the directors then in
office, though less than a quorum and whether or not elected by the same class or series of stock as is entitled to vote for the election
of the directors to fill such vacancy, or by a sole remaining director, and the directors so chosen shall hold office until the next annual
election and until their successors are duly elected and shall qualify, unless sooner displaced; provided, however, that where such vacancy
occurs among the directors elected by the holders of a class or series of stock, the holders of shares of such class or series may override
the Board’s action to fill such vacancy by removing any person so appointed by the Board at any time and electing their own designee.
3.3
Preferred Stock Protective Provisions. At any time when at least 5,253,332 shares of Preferred Stock (subject to appropriate
adjustment in the event of any stock dividend, stock split, combination or other similar recapitalization with respect to the Preferred
Stock) are outstanding, the Corporation shall not, either directly or indirectly by amendment, merger, consolidation, domestication, transfer,
continuance, recapitalization, reclassification, waiver, statutory conversion or otherwise, do any of the following without (in addition
to any other vote required by law or this Amended and Restated Certificate of Incorporation) the written consent or affirmative vote of
the Requisite Holders given in writing or by vote at a meeting, consenting or voting (as the case may be) separately as a class, and any
such act or transaction entered into without such consent or vote shall be null and void ab initio, and of no force or effect;
provided, however, that a consent or resolution of the Board of Directors approving any of the following items that
is contingent upon receipt of the stockholder consent required pursuant to this Section 3.3 shall not, solely by reason thereof,
be deemed to be an action or transaction entered into in violation hereof.
3.3.1
liquidate, dissolve or wind-up the business and affairs of the Corporation or effect any merger or consolidation or any other Deemed
Liquidation Event, or consent to any of the foregoing;
3.3.2
amend, alter or repeal any provision of this Amended and Restated Certificate of Incorporation or Bylaws of the Corporation;
3.3.3
(i) create, or authorize the creation of, or issue or obligate itself to issue shares of, or reclassify, any capital stock, or
create or issue or obligate itself to issue any other security convertible into or exercisable for any equity security, unless the same
ranks junior to the Preferred Stock with respect to its rights, preferences and privileges, or (ii) increase the authorized number of
shares of Preferred Stock or any additional class or series of capital stock of the Corporation;
3.3.4
purchase or redeem (or permit any subsidiary to purchase or redeem) or pay or declare any dividend or make any distribution on,
any shares of capital stock of the Corporation other than (i) redemptions of or dividends or distributions on the Preferred Stock as expressly
authorized herein, (ii) dividends or other distributions payable on the Common Stock solely in the form of additional shares of Common
Stock and (iii) repurchases of stock from employees, officers, directors, consultants or other persons who performed services for
the Corporation or any subsidiary upon the occurrence of certain events, such as (i) in connection with the cessation of such employment
or service at no greater than the original purchase price thereof or (ii) pursuant to a right of first refusal or (iii) as approved by
the Board of Directors;
3.3.5
create, adopt, amend, terminate or repeal any equity (or equity-linked) compensation plan;
3.3.6
create, or authorize the creation of, or issue, or authorize the issuance of any debt security or create any lien or security interest
(except for purchase money liens or statutory liens of landlords, mechanics, materialmen, workmen, warehousemen and other similar persons
arising or incurred in the ordinary course of business) or incur other indebtedness for borrowed money, including but not limited to obligations
and contingent obligations under guarantees, or permit any subsidiary to take any such action with respect to any debt security lien,
security interest or other indebtedness for borrowed money, other than equipment leases, bank lines of credit or trade payables incurred
in the ordinary course or unless approved by the Board of Directors, including a majority of the Preferred Directors then seated; or
3.3.7
increase or decrease the size of the Board of Directors;
3.3.8
create, or hold capital stock in, any subsidiary that is not wholly owned (either directly or through one (1) or more other subsidiaries)
by the Corporation, or permit any subsidiary to create, or authorize the creation of, or issue or obligate itself to issue, any shares
of any class or series of capital stock, or sell, transfer or otherwise dispose of any capital stock of any direct or indirect subsidiary
of the Corporation, or permit any direct or indirect subsidiary to sell, lease, transfer, exclusively license or otherwise dispose (in
a single transaction or series of related transactions) of all or substantially all of the assets of such subsidiary.
3.4
Series B Preferred Stock Protective Provisions. At any time when at least 1,545,070 shares of Series B Preferred Stock and
Series B-1 Preferred Stock in the aggregate (in each case subject to appropriate adjustment in the event of any stock dividend, stock
split, combination or other similar recapitalization with respect to the Preferred Stock) are outstanding, the Corporation shall not,
either directly or indirectly by amendment, merger, consolidation, domestication, transfer, continuance, recapitalization, reclassification,
waiver, statutory conversion or otherwise, do any of the following without (in addition to any other vote required by law or this Amended
and Restated Certificate of Incorporation) the written consent or affirmative vote of the holders of at least 56% of the outstanding shares
of Series B Preferred Stock and Series B-1 Preferred Stock, voting together as a separate class on an as-converted basis, which must include
the written consent or affirmative vote of at least one of the Lead Purchasers (as defined in the Voting Agreement) (the “Series
B Requisite Holders”), given in writing or by vote at a meeting, consenting or voting (as the case may be) separately as a class,
and any such act or transaction entered into without such consent or vote shall be null and void ab initio, and of no force
or effect; provided, however, that a consent or resolution of the Board of Directors approving any of the following
items that is contingent upon receipt of the stockholder consent required pursuant to this Section 3.4 shall not, solely by reason
thereof, be deemed to be an action or transaction entered into in violation hereof.
3.4.1
modify, amend or waive the rights, preferences or privileges of the Preferred Stock, the Series B Preferred Stock or the Series
B-1 Preferred Stock so as to affect adversely any rights, preferences or privileges expressly and uniquely granted to the Series B Preferred
Stock or Series B-1 Preferred Stock; provided, however, that the authorization of one or more additional series of Preferred Stock with
a liquidation preference and/or other rights, preferences or privileges on parity with or senior to the Series B Preferred Stock or Series
B-1 Preferred Stock, as the case may be, pursuant to a bona fide financing transaction shall not be deemed to affect adversely the rights,
preferences or privileges of the Series B Preferred Stock or Series B-1 Preferred Stock;
3.4.2
increase or decrease the number of authorized shares of Series B Preferred Stock or Series B-1 Preferred Stock; or
3.4.3
purchase or redeem or pay any dividend on any capital stock other than stock repurchased from former employees and consultants
in connection with the cessation of their service at the lower of fair market value or the original purchase price thereof.
3.5
Series A Preferred Stock Protective Provisions. At any time when at least 3,524,006 shares of Series A Preferred Stock (subject
to appropriate adjustment in the event of any stock dividend, stock split, combination or other similar recapitalization with respect
to the Series A Preferred Stock) are outstanding the Corporation shall not, either directly or indirectly by amendment, merger, consolidation,
domestication, transfer, continuance, recapitalization, reclassification, waiver, statutory conversion or otherwise, do any of the following
without (in addition to any other vote required by law or this Amended and Restated Certificate of Incorporation) the written consent
or affirmative vote of the holders of a majority of the outstanding shares of Series A Preferred Stock given in writing or by vote at
a meeting, consenting or voting (as the case may be) separately as a class, and any such act or transaction entered into without
such consent or vote shall be null and void ab initio, and of no force or effect; provided, however, that
a consent or resolution of the Board of Directors approving any of the following items that is contingent upon receipt of the stockholder
consent required pursuant to this Section 3.4 shall not, solely by reason thereof, be deemed to be an action or transaction entered
into in violation hereof.
3.5.1
modify, amend or waive the rights, preferences or privileges of the Preferred Stock or the Series A Preferred Stock so as to affect
adversely any rights, preferences or privileges expressly and uniquely granted to the Series A Preferred Stock; provided, however, that
the authorization of one or more additional series of Preferred Stock with a liquidation preference and/or other rights, preferences or
privileges on parity with or senior to the Series A Preferred Stock pursuant to a bona fide financing transaction shall not be deemed
to affect adversely the rights, preferences or privileges of the Series A Preferred Stock;
3.5.2
increase or decrease the number of authorized shares of Series A Preferred Stock; or
3.5.3
purchase or redeem or pay any dividend on any capital stock other than stock repurchased from former employees and consultants
in connection with the cessation of their service at the lower of fair market value or the original purchase price thereof.
4.
Optional Conversion. The holders of the Preferred Stock shall have conversion rights as follows (the “Conversion
Rights”):
4.1
Right to Convert.
4.1.1
Conversion Ratio. Each share of Preferred Stock shall be convertible, at the option of the holder thereof, at any time and
from time to time, and without the payment of additional consideration by the holder thereof, into such number of fully paid and non-assessable
shares of Common Stock as is determined by dividing the applicable Original Issue Price by the applicable Conversion Price (as defined
below) in effect at the time of conversion. The “Conversion Price” (i) applicable to the Series Seed Preferred Stock
shall initially be equal to $1.0000, (ii) applicable to the Series A Preferred Stock shall initially be equal to $10.0295, (iii) applicable
to the Series B-1 Preferred Stock shall initially be equal to $12.4351 and (iv) applicable to the Series B Preferred Stock shall initially
be equal to $14.6295. Such initial Conversion Price, and the rate at which shares of Preferred Stock may be converted into shares of Common
Stock, shall be subject to adjustment as provided below.
4.1.2
Termination of Conversion Rights. In the event of a liquidation, dissolution or winding up of the Corporation or a Deemed
Liquidation Event, the Conversion Rights shall terminate at the close of business on the last full day preceding the date fixed for the
payment of any such amounts distributable on such event to the holders of Preferred Stock; provided that the foregoing termination
of Conversion Rights shall not affect the amount(s) otherwise paid or payable in accordance with Section 2.1 to holders of Preferred
Stock pursuant to such liquidation, dissolution or winding up of the Corporation or a Deemed Liquidation Event.
4.2
Fractional Shares. No fractional shares of Common Stock shall be issued upon conversion of the Preferred Stock. In lieu
of any fractional shares to which the holder would otherwise be entitled, the number of shares of Common Stock to be issued upon conversion
of the Preferred Stock shall be rounded to the nearest whole share.
4.3
Mechanics of Conversion.
4.3.1
Notice of Conversion. In order for a holder of Preferred Stock to voluntarily convert shares of Preferred Stock into shares
of Common Stock, such holder shall (a) provide written notice to the Corporation’s transfer agent at the office of the transfer
agent for the Preferred Stock (or at the principal office of the Corporation if the Corporation serves as its own transfer agent) that
such holder elects to convert all or any number of such holder’s shares of Preferred Stock and, if applicable, any event on which
such conversion is contingent and (b), if such holder’s shares are certificated, surrender the certificate or certificates for such
shares of Preferred Stock (or, if such registered holder alleges that such certificate has been lost, stolen or destroyed, the Lost Certificate
Documentation), at the office of the transfer agent for the Preferred Stock (or at the principal office of the Corporation if the Corporation
serves as its own transfer agent). Such notice shall state such holder’s name or the names of the nominees in which such holder
wishes the shares of Common Stock to be issued. If required by the Corporation, any certificates surrendered for conversion shall be endorsed
or accompanied by a written instrument or instruments of transfer, in form satisfactory to the Corporation, duly executed by the registered
holder or his, her or its attorney duly authorized in writing. The close of business on the date of receipt by the transfer agent (or
by the Corporation if the Corporation serves as its own transfer agent) of such notice and, if applicable, certificates (or the Lost Certificate
Documentation) shall be the time of conversion (the “Conversion Time”), and the shares of Common Stock issuable upon
conversion of the specified shares shall be deemed to be outstanding of record as of such date. The Corporation shall, as soon as practicable
after the Conversion Time (i) issue and deliver to such holder of Preferred Stock, or to his, her or its nominees, a notice of issuance
of uncertificated shares and may, upon written request, issue and deliver a certificate for the number of full shares of Common Stock
issuable upon such conversion in accordance with the provisions hereof and, may, if applicable and upon written request, issue and deliver
a certificate for the number (if any) of the shares of Preferred Stock represented by any surrendered certificate that were not converted
into Common Stock and (ii) pay all declared but unpaid dividends on the shares of Preferred Stock converted.
4.3.2
Reservation of Shares. The Corporation shall at all times when the Preferred Stock shall be outstanding, reserve and keep
available out of its authorized but unissued capital stock, for the purpose of effecting the conversion of the Preferred Stock, such number
of its duly authorized shares of Common Stock as shall from time to time be sufficient to effect the conversion of all outstanding Preferred
Stock; and if at any time the number of authorized but unissued shares of Common Stock shall not be sufficient to effect the conversion
of all then outstanding shares of the Preferred Stock, the Corporation shall take such corporate action as may be necessary to increase
its authorized but unissued shares of Common Stock to such number of shares as shall be sufficient for such purposes, including, without
limitation, engaging in best efforts to obtain the requisite stockholder approval of any necessary amendment to this Amended and Restated
Certificate of Incorporation. Before taking any action which would cause an adjustment reducing the Conversion Price below the then par
value of the shares of Common Stock issuable upon conversion of the Preferred Stock, the Corporation will take any corporate action which
may, in the opinion of its counsel, be necessary in order that the Corporation may validly and legally issue fully paid and non-assessable
shares of Common Stock at such adjusted Conversion Price.
4.3.3
Effect of Conversion. All shares of Preferred Stock which shall have been surrendered for conversion as herein provided
shall no longer be deemed to be outstanding and all rights with respect to such shares shall immediately cease and terminate at the Conversion
Time, except only the right of the holders thereof to receive shares of Common Stock in exchange therefor and to receive payment of any
dividends declared but unpaid thereon. Any shares of Preferred Stock so converted shall be retired and cancelled and may not be reissued
as shares of such series, and the Corporation may thereafter take such appropriate action (without the need for stockholder action) as
may be necessary to reduce the authorized number of shares of Preferred Stock accordingly.
4.3.4
No Further Adjustment. Upon any such conversion, no adjustment to the Conversion Price shall be made for any declared but
unpaid dividends on the Preferred Stock surrendered for conversion or on the Common Stock delivered upon conversion.
4.3.5
Taxes. The Corporation shall pay any and all issue and other similar taxes that may be payable in respect of any issuance
or delivery of shares of Common Stock upon conversion of shares of Preferred Stock pursuant to this Section 4. The Corporation
shall not, however, be required to pay any tax which may be payable in respect of any transfer involved in the issuance and delivery of
shares of Common Stock in a name other than that in which the shares of Preferred Stock so converted were registered, and no such issuance
or delivery shall be made unless and until the person or entity requesting such issuance has paid to the Corporation the amount of any
such tax or has established, to the satisfaction of the Corporation, that such tax has been paid.
4.4
Adjustments to Conversion Price for Diluting Issues.
4.4.1
Special Definitions. For purposes of this Article Fourth, the following definitions shall apply:
(a)
“Additional Shares of Common Stock” shall mean all shares of Common Stock issued (or, pursuant to Section 4.4.3
below, deemed to be issued) by the Corporation after the date upon which the first share of Series B Preferred Stock is issued (the “Original
Issue Date”), other than (1) the following shares of Common Stock and (2) shares of Common Stock deemed issued pursuant to the
following Options (as defined below) and Convertible Securities (as defined below) (clauses (1) and (2), collectively, “Exempted
Securities”):
| (i) | as to any series of Preferred Stock shares of Common Stock, Options or Convertible Securities issued as
a dividend or distribution on such series of Preferred Stock; |
| (ii) | shares of Common Stock, Options or Convertible Securities issued by reason of a dividend, stock split,
split-up or other distribution on shares of Common Stock that is covered by Section 4.5, 4.6, 4.7 or 4.8; |
| (iii) | shares of Common Stock or Options issued to employees or directors of, or consultants or advisors to,
the Corporation or any of its subsidiaries pursuant to a plan, agreement or arrangement approved by the Board of Directors; |
| (iv) | shares of Common Stock or Convertible Securities actually issued upon the exercise of Options or shares
of Common Stock actually issued upon the conversion or exchange of Convertible Securities, in each case provided such issuance is pursuant
to the terms of such Option or Convertible Security; |
| (v) | shares of Common Stock, Options or Convertible Securities issued to banks, equipment lessors or other
financial institutions, or to real property lessors, pursuant to a debt financing, equipment leasing or real property leasing transaction
approved by the Board of Directors; |
| (vi) | shares of Common Stock, Options or Convertible Securities issued to suppliers or third party service providers
in connection with the provision of goods or services pursuant to transactions approved by the Board of Directors; |
| (vii) | shares of Common Stock, Options or Convertible Securities issued as acquisition consideration pursuant
to the acquisition of another corporation by the Corporation by merger, purchase of substantially all of the assets or other reorganization
or to a joint venture agreement, provided that such issuances are approved by the Board of Directors; |
| (viii) | shares of Common Stock, Options or Convertible Securities issued in connection with sponsored research,
collaboration, technology license, development, OEM, marketing or other similar agreements or strategic partnerships approved by the Board
of Directors; or |
| (ix) | shares of Common Stock issued in connection with a Qualified IPO (as defined below). |
(b) “Convertible Securities” shall mean any evidences of indebtedness, shares or other securities directly or indirectly
convertible into or exchangeable for Common Stock, but excluding Options.
(c) “Option” shall mean rights, options or warrants to subscribe for, purchase or otherwise acquire Common Stock
or Convertible Securities.
4.4.2
No Adjustment of Conversion Price. No adjustment in the Conversion Price shall be made as the result of the issuance or
deemed issuance of Additional Shares of Common Stock if the Corporation receives written notice from the Requisite Holders agreeing that
no such adjustment shall be made as the result of the issuance or deemed issuance of such Additional Shares of Common Stock. In addition,
(i) no adjustment in the Conversion Price applicable to the Series A Preferred Stock shall be made as the result of the issuance or deemed
issuance of Additional Shares of Common Stock if the Corporation receives written notice from the holders of a majority of the outstanding
shares of Series A Preferred Stock agreeing that no such adjustment shall be made as the result of the issuance or deemed issuance of
such Additional Shares of Common Stock and (ii) no adjustment in the Conversion Price applicable to the Series B Preferred Stock shall
be made as the result of the issuance or deemed issuance of Additional Shares of Common Stock if the Corporation receives written notice
from the Series B Requisite Holders agreeing that no such adjustment shall be made as the result of the issuance or deemed issuance of
such Additional Shares of Common Stock.
4.4.3
Deemed Issue of Additional Shares of Common Stock.
(a)
If the Corporation at any time or from time to time after the Original Issue Date shall issue any Options or Convertible Securities
(excluding Options or Convertible Securities which are themselves Exempted Securities) or shall fix a record date for the determination
of holders of any class of securities entitled to receive any such Options or Convertible Securities, then the maximum number of shares
of Common Stock (as set forth in the instrument relating thereto, assuming the satisfaction of any conditions to exercisability, convertibility
or exchangeability but without regard to any provision contained therein for a subsequent adjustment of such number) issuable upon the
exercise of such Options or, in the case of Convertible Securities and Options therefor, the conversion or exchange of such Convertible
Securities, shall be deemed to be Additional Shares of Common Stock issued as of the time of such issue or, in case such a record date
shall have been fixed, as of the close of business on such record date.
(b)
If the terms of any Option or Convertible Security, the issuance of which resulted in an adjustment to the Conversion Price pursuant
to the terms of Section 4.4.4, are revised as a result of an amendment to such terms or any other adjustment pursuant to the provisions
of such Option or Convertible Security (but excluding automatic adjustments to such terms pursuant to anti-dilution or similar provisions
of such Option or Convertible Security) to provide for either (1) any increase or decrease in the number of shares of Common Stock issuable
upon the exercise, conversion and/or exchange of any such Option or Convertible Security or (2) any increase or decrease in the consideration
payable to the Corporation upon such exercise, conversion and/or exchange, then, effective upon such increase or decrease becoming effective,
the Conversion Price computed upon the original issue of such Option or Convertible Security (or upon the occurrence of a record date
with respect thereto) shall be readjusted to such Conversion Price as would have obtained had such revised terms been in effect upon the
original date of issuance of such Option or Convertible Security. Notwithstanding the foregoing, no readjustment pursuant to this clause (b)
shall have the effect of increasing the Conversion Price to an amount which exceeds the lower of (i) the Conversion Price in effect immediately
prior to the original adjustment made as a result of the issuance of such Option or Convertible Security, or (ii) the Conversion Price
that would have resulted from any issuances of Additional Shares of Common Stock (other than deemed issuances of Additional Shares of
Common Stock as a result of the issuance of such Option or Convertible Security) between the original adjustment date and such readjustment
date.
(c)
If the terms of any Option or Convertible Security (excluding Options or Convertible Securities which are themselves Exempted Securities),
the issuance of which did not result in an adjustment to the Conversion Price pursuant to the terms of Section 4.4.4 (either because
the consideration per share (determined pursuant to Section 4.4.5) of the Additional Shares of Common Stock subject thereto was
equal to or greater than the Conversion Price then in effect, or because such Option or Convertible Security was issued before the Original
Issue Date), are revised after the Original Issue Date as a result of an amendment to such terms or any other adjustment pursuant to the
provisions of such Option or Convertible Security (but excluding automatic adjustments to such terms pursuant to anti-dilution or similar
provisions of such Option or Convertible Security) to provide for either (1) any increase in the number of shares of Common Stock issuable
upon the exercise, conversion or exchange of any such Option or Convertible Security or (2) any decrease in the consideration payable
to the Corporation upon such exercise, conversion or exchange, then such Option or Convertible Security, as so amended or adjusted, and
the Additional Shares of Common Stock subject thereto (determined in the manner provided in Section 4.4.3(a) shall be deemed to
have been issued effective upon such increase or decrease becoming effective).
(d)
Upon the expiration or termination of any unexercised Option or unconverted or unexchanged Convertible Security (or portion thereof)
which resulted (either upon its original issuance or upon a revision of its terms) in an adjustment to the Conversion Price pursuant to
the terms of Section 4.4.4, the Conversion Price shall be readjusted to such Conversion Price as would have obtained had such
Option or Convertible Security (or portion thereof) never been issued.
(e)
If the number of shares of Common Stock issuable upon the exercise, conversion and/or exchange of any Option or Convertible Security,
or the consideration payable to the Corporation upon such exercise, conversion and/or exchange, is calculable at the time such Option
or Convertible Security is issued or amended but is subject to adjustment based upon subsequent events, any adjustment to the Conversion
Price provided for in this Section 4.4.3 shall be effected at the time of such issuance or amendment based on such number
of shares or amount of consideration without regard to any provisions for subsequent adjustments (and any subsequent adjustments shall
be treated as provided in clauses (b) and (c) of this Section 4.4.3). If the number of shares of Common Stock
issuable upon the exercise, conversion and/or exchange of any Option or Convertible Security, or the consideration payable to the Corporation
upon such exercise, conversion and/or exchange, cannot be calculated at all at the time such Option or Convertible Security is issued
or amended, any adjustment to the Conversion Price that would result under the terms of this Section 4.4.3 at the time of
such issuance or amendment shall instead be effected at the time such number of shares and/or amount of consideration is first calculable
(even if subject to subsequent adjustments), assuming for purposes of calculating such adjustment to the Conversion Price that such issuance
or amendment took place at the time such calculation can first be made.
4.4.4 Adjustment of Conversion Price Upon Issuance of Additional Shares of Common Stock. In the event the Corporation shall at
any time after the Original Issue Date issue Additional Shares of Common Stock (including Additional Shares of Common Stock deemed to
be issued pursuant to Section 4.4.3), without consideration or for a consideration per share less than the Conversion Price
in effect immediately prior to such issuance or deemed issuance, then the Conversion Price shall be reduced, concurrently with such issue,
to a price (calculated to the nearest one-hundredth of a cent) determined in accordance with the following formula:
CP2 = CP1* (A + B) ÷
(A + C).
For purposes of the foregoing formula, the following
definitions shall apply:
(a)
“CP2” shall mean the Conversion Price in effect immediately after such issuance or deemed issuance of Additional
Shares of Common Stock
(b)
“CP1” shall mean the Conversion Price in effect immediately prior to such issuance or deemed issuance of
Additional Shares of Common Stock;
(c)
“A” shall mean the number of shares of Common Stock outstanding immediately prior to such issuance or deemed issuance
of Additional Shares of Common Stock (treating for this purpose as outstanding all shares of Common Stock issuable upon exercise of Options
outstanding immediately prior to such issuance or deemed issuance or upon conversion or exchange of Convertible Securities (including
the Preferred Stock) outstanding (assuming exercise of any outstanding Options therefor) immediately prior to such issue);
(d)
“B” shall mean the number of shares of Common Stock that would have been issued if such Additional Shares of Common
Stock had been issued or deemed issued at a price per share equal to CP1 (determined by dividing the aggregate consideration
received by the Corporation in respect of such issue by CP1); and
(e)
“C” shall mean the number of such Additional Shares of Common Stock issued in such transaction.
4.4.5
Determination of Consideration. For purposes of this Section 4.4, the consideration received by the Corporation for
the issuance or deemed issuance of any Additional Shares of Common Stock shall be computed as follows:
(a)
Cash and Property. Such consideration shall:
| (i) | insofar as it consists of cash, be computed at the aggregate amount of cash received by the Corporation,
excluding amounts paid or payable for accrued interest; |
| (ii) | insofar as it consists of property other than cash, be computed at the fair market value thereof at the
time of such issue, as determined in good faith by the Board of Directors, including a majority of the Preferred Directors then seated;
and |
| (iii) | in the event Additional Shares of Common Stock are issued together with other shares or securities or
other assets of the Corporation for consideration which covers both, be the proportion of such consideration so received, computed as
provided in clauses (i) and (ii) above, as determined in good faith by the Board of Directors, including a majority of the
Preferred Directors (as defined below) then seated. |
(b)
Options and Convertible Securities. The consideration per share received by the Corporation for Additional Shares of Common
Stock deemed to have been issued pursuant to Section 4.4.3, relating to Options and Convertible Securities, shall be determined
by dividing:
| (i) | The total amount, if any, received or receivable by the Corporation as consideration for the issue of
such Options or Convertible Securities, plus the minimum aggregate amount of additional consideration (as set forth in the instruments
relating thereto, without regard to any provision contained therein for a subsequent adjustment of such consideration) payable to the
Corporation upon the exercise of such Options or the conversion or exchange of such Convertible Securities, or in the case of Options
for Convertible Securities, the exercise of such Options for Convertible Securities and the conversion or exchange of such Convertible
Securities, by |
| (ii) | the maximum number of shares of Common Stock (as set forth in the instruments relating thereto, without
regard to any provision contained therein for a subsequent adjustment of such number) issuable upon the exercise of such Options or the
conversion or exchange of such Convertible Securities, or in the case of Options for Convertible Securities, the exercise of such Options
for Convertible Securities and the conversion or exchange of such Convertible Securities. |
4.4.6
Multiple Closing Dates. In the event the Corporation shall issue on more than one date Additional Shares of Common Stock
that are a part of one transaction or a series of related transactions and that would result in an adjustment to the Conversion Price
pursuant to the terms of Section 4.4.4, and such issuance dates occur within a period of no more than 90 days from the first such
issuance to the final such issuance, then, upon the final such issuance, the Conversion Price shall be readjusted to give effect to all
such issuances as if they occurred on the date of the first such issuance (and without giving effect to any additional adjustments as
a result of any such subsequent issuances within such period).
4.5
Adjustment for Stock Splits and Combinations. If the Corporation shall at any time or from time to time after the Original
Issue Date effect a subdivision of the outstanding shares of Common Stock, the Conversion Price in effect immediately before that subdivision
shall be proportionately decreased so that the number of shares of Common Stock issuable on conversion of each share of such series shall
be increased in proportion to such increase in the aggregate number of shares of Common Stock outstanding. If the Corporation shall at
any time or from time to time after the Original Issue Date combine the outstanding shares of Common Stock, the Conversion Price in effect
immediately before the combination shall be proportionately increased so that the number of shares of Common Stock issuable on conversion
of each share of such series shall be decreased in proportion to such decrease in the aggregate number of shares of Common Stock outstanding.
Any adjustment under this Section shall become effective at the close of business on the date the subdivision or combination becomes effective.
4.6
Adjustment for Certain Dividends and Distributions. In the event the Corporation at any time or from time to time after
the Original Issue Date shall make or issue, or fix a record date for the determination of holders of Common Stock entitled to receive,
a dividend or other distribution payable on the Common Stock in additional shares of Common Stock, then and in each such event the Conversion
Price in effect immediately before such event shall be decreased as of the time of such issuance or, in the event such a record date shall
have been fixed, as of the close of business on such record date, by multiplying the Conversion Price then in effect by a fraction:
(1)
the numerator of which shall be the total number of shares of Common Stock issued and outstanding immediately prior to the time
of such issuance or the close of business on such record date, and
(2)
the denominator of which shall be the total number of shares of Common Stock issued and outstanding immediately prior to the time
of such issuance or the close of business on such record date plus the number of shares of Common Stock issuable in payment of such dividend
or distribution.
Notwithstanding the foregoing, (a) if such record
date shall have been fixed and such dividend is not fully paid or if such distribution is not fully made on the date fixed therefor, the
Conversion Price shall be recomputed accordingly as of the close of business on such record date and thereafter the Conversion Price shall
be adjusted pursuant to this Section as of the time of actual payment of such dividends or distributions; and (b) that no such adjustment
shall be made if the holders of Preferred Stock simultaneously receive a dividend or other distribution of shares of Common Stock in a
number equal to the number of shares of Common Stock as they would have received if all outstanding shares of Preferred Stock had been
converted into Common Stock on the date of such event.
4.7
Adjustments for Other Dividends and Distributions. In the event the Corporation at any time or from time to time after the
Original Issue Date shall make or issue, or fix a record date for the determination of holders of Common Stock entitled to receive, a
dividend or other distribution payable in securities of the Corporation (other than a distribution of shares of Common Stock in respect
of outstanding shares of Common Stock) or in other property and the provisions of Section 1 do not apply to such dividend
or distribution, then and in each such event the holders of Preferred Stock shall receive, simultaneously with the distribution to the
holders of Common Stock, a dividend or other distribution of such securities or other property in an amount equal to the amount of such
securities or other property as they would have received if all outstanding shares of Preferred Stock had been converted into Common Stock
on the date of such event.
4.8
Adjustment for Merger or Reorganization, etc. Subject to the provisions of Section 2.3, if there shall occur any
reorganization, recapitalization, reclassification, consolidation or merger involving the Corporation in which the Common Stock (but not
the Preferred Stock) is converted into or exchanged for securities, cash or other property (other than a transaction covered by Sections
4.4, 4.6 or 4.7), then, following any such reorganization, recapitalization, reclassification, consolidation or merger,
each share of Preferred Stock shall thereafter be convertible in lieu of the Common Stock into which it was convertible prior to such
event into the kind and amount of securities, cash or other property which a holder of the number of shares of Common Stock of the Corporation
issuable upon conversion of one share of Preferred Stock immediately prior to such reorganization, recapitalization, reclassification,
consolidation or merger would have been entitled to receive pursuant to such transaction; and, in such case, appropriate adjustment (as
determined in good faith by the Board of Directors) shall be made in the application of the provisions in this Section 4 with respect
to the rights and interests thereafter of the holders of the Preferred Stock, to the end that the provisions set forth in this Section
4 (including provisions with respect to changes in and other adjustments of the Conversion Price) shall thereafter be applicable,
as nearly as reasonably may be, in relation to any securities or other property thereafter deliverable upon the conversion of the Preferred
Stock.
4.9
Certificate as to Adjustments. Upon the occurrence of each adjustment or readjustment of the Conversion Price pursuant to
this Section 4, the Corporation at its expense shall, as promptly as reasonably practicable but in any event not later than ten
days thereafter, compute such adjustment or readjustment in accordance with the terms hereof and furnish to each holder of Preferred Stock
a certificate setting forth such adjustment or readjustment (including the kind and amount of securities, cash or other property into
which the Preferred Stock is convertible) and showing in detail the facts upon which such adjustment or readjustment is based. The Corporation
shall, as promptly as reasonably practicable after the written request at any time of any holder of Preferred Stock (but in any event
not later than ten days thereafter), furnish or cause to be furnished to such holder a certificate setting forth (i) the Conversion Price
then in effect, and (ii) the number of shares of Common Stock and the amount, if any, of other securities, cash or property which then
would be received upon the conversion of Preferred Stock.
4.10
Notice of Record Date. In the event:
(a)
the Corporation shall take a record of the holders of its Common Stock (or other capital stock or securities at the time issuable
upon conversion of the Preferred Stock) for the purpose of entitling or enabling them to receive any dividend or other distribution, or
to receive any right to subscribe for or purchase any shares of capital stock of any class or any other securities, or to receive any
other security; or
(b)
of any capital reorganization of the Corporation, any reclassification of the Common Stock of the Corporation, or any Deemed Liquidation
Event; or
(c)
of the voluntary or involuntary dissolution, liquidation or winding-up of the Corporation,
then, and in each such case, the Corporation will
send or cause to be sent to the holders of the Preferred Stock a notice specifying, as the case may be, (i) the record date for such dividend,
distribution or right, and the amount and character of such dividend, distribution or right, or (ii) the effective date on which such
reorganization, reclassification, consolidation, merger, transfer, dissolution, liquidation or winding-up is proposed to take place, and
the time, if any is to be fixed, as of which the holders of record of Common Stock (or such other capital stock or securities at the time
issuable upon the conversion of the Preferred Stock) shall be entitled to exchange their shares of Common Stock (or such other capital
stock or securities) for securities or other property deliverable upon such reorganization, reclassification, consolidation, merger, transfer,
dissolution, liquidation or winding-up, and the amount per share and character of such exchange applicable to the Preferred Stock and
the Common Stock. Such notice shall be sent at least ten days prior to the record date or effective date for the event specified in such
notice.
5.
Mandatory Conversion.
5.1
Trigger Events. Upon either (a) the closing of the sale of shares of Common Stock to the public in a firm-commitment underwritten
public offering pursuant to an effective registration statement under the Securities Act of 1933, as amended, resulting in at least $75,000,000
of gross proceeds to the Corporation and in connection with such offering the Common Stock is listed for trading on the Nasdaq Stock Market’s
National Market, the New York Stock Exchange or another exchange or marketplace approved by the Board of Directors (a “Qualified
IPO”), or (b) the date and time, or the occurrence of an event, specified by vote or written consent of the Requisite Holders
(the time of such closing or the date and time specified or the time of the event specified in such vote or written consent is referred
to herein as the “Mandatory Conversion Time”), then (i) all outstanding shares of Preferred Stock shall automatically
be converted into shares of Common Stock, at the then effective conversion rate as calculated pursuant to Section 4.1.1 and (ii)
such shares may not be reissued by the Corporation.
5.2
Procedural Requirements. All holders of record of shares of Preferred Stock shall be sent written notice of the Mandatory
Conversion Time and the place designated for mandatory conversion of all such shares of Preferred Stock pursuant to this Section 5.
Such notice need not be sent in advance of the occurrence of the Mandatory Conversion Time. Upon receipt of such notice, each holder of
shares of Preferred Stock in certificated form shall surrender his, her or its certificate or certificates for all such shares (or, if
such holder alleges that such certificate has been lost, stolen or destroyed, the Lost Certificate Documentation) to the Corporation at
the place designated in such notice. If so required by the Corporation, any certificates surrendered for conversion shall be endorsed
or accompanied by written instrument or instruments of transfer, in form satisfactory to the Corporation, duly executed by the registered
holder or by his, her or its attorney duly authorized in writing. All rights with respect to the Preferred Stock converted pursuant to
Section 5.1, including the rights, if any, to receive notices and vote (other than as a holder of Common Stock), will terminate
at the Mandatory Conversion Time (notwithstanding the failure of the holder or holders thereof to surrender any certificates at or prior
to such time), except only the rights of the holders thereof, upon surrender of any certificate or certificates of such holders (or the
Lost Certificate Documentation) therefor, to receive the items provided for in the next sentence of this Section 5.2. As soon as
practicable after the Mandatory Conversion Time and, if applicable, the surrender of any certificate or certificates (or the Lost Certificate
Documentation) for Preferred Stock, the Corporation shall (a) issue and deliver to such holder, or to his, her or its nominees, a notice
of issuance of uncertificated shares and may, upon written request, issue and deliver a certificate for the number of full shares of Common
Stock issuable upon such conversion in accordance with the provisions hereof and (b) pay any declared but unpaid dividends on the shares
of Preferred Stock converted. Such converted Preferred Stock shall be retired and cancelled and may not be reissued as shares of such
series, and the Corporation may thereafter take such appropriate action (without the need for stockholder action) as may be necessary
to reduce the authorized number of shares of Preferred Stock accordingly.
6.
Redeemed or Otherwise Acquired Shares. Any shares of Preferred Stock that are redeemed, converted or otherwise acquired
by the Corporation or any of its subsidiaries shall be automatically and immediately cancelled and retired and shall not be reissued,
sold or transferred. Neither the Corporation nor any of its subsidiaries may exercise any voting or other rights granted to the holders
of Preferred Stock following redemption, conversion or acquisition.
7.
Waiver. Except as otherwise set forth herein, (a) any of the rights, powers, preferences and other terms of the Preferred
Stock set forth herein may be waived on behalf of all holders of Preferred Stock by the affirmative written consent or vote of the holders
that would otherwise be required to amend such right, powers, preferences, and other terms and (b) at any time more than one series of
Preferred Stock is issued and outstanding, any of the rights, powers, preferences and other terms of any series of Preferred Stock set
forth herein may be waived on behalf of all holders of such series of Preferred Stock by the affirmative written consent or vote of the
holders of such series that would otherwise be required to amend such right, power, preference, or other term.
8.
Notices. Any notice required or permitted by the provisions of this Article Fourth to be given to a holder of shares of
Preferred Stock shall be mailed, postage prepaid, to the post office address last shown on the records of the Corporation, or given by
electronic communication in compliance with the provisions of the General Corporation Law, and shall be deemed sent upon such mailing
or electronic transmission.
Fifth:
Subject to any additional vote required by this Amended and Restated Certificate of Incorporation or Bylaws, in furtherance and not in
limitation of the powers conferred by statute, the Board of Directors is expressly authorized to make, repeal, alter, amend and rescind
any or all of the Bylaws of the Corporation.
Sixth:
Subject to any additional vote required by this Amended and Restated Certificate of Incorporation, the number of directors of the Corporation
shall be determined in the manner set forth in the Bylaws of the Corporation. Each director shall be entitled to one vote on each matter
presented to the Board of Directors.
Seventh:
Elections of directors need not be by written ballot unless the Bylaws of the Corporation shall so provide.
Eighth:
Meetings of stockholders may be held within or without the State of Delaware, as the Bylaws of the Corporation may provide. The books
of the Corporation may be kept outside the State of Delaware at such place or places as may be designated from time to time by the Board
of Directors or in the Bylaws of the Corporation.
Ninth:
To the fullest extent permitted by law, a director or officer of the Corporation shall not be personally liable to the Corporation or
its stockholders for monetary damages for breach of fiduciary duty as a director or officer. If the General Corporation Law or any other
law of the State of Delaware is amended after approval by the stockholders of this Article Ninth to authorize corporate action further
eliminating or limiting the personal liability of directors or officers, then the liability of a director or officer of the Corporation
shall be eliminated or limited to the fullest extent permitted by the General Corporation Law as so amended. Solely for the purposes of
this Article NINTH, “officer” shall have the meaning provided in Section 102(b)(7) of the General Corporation Law as amended
from time to time.
Any
amendment, repeal or modification of the foregoing provisions of this Article Ninth by the stockholders of the Corporation shall
not adversely affect any right or protection of a director or officer of the Corporation existing at the time of, or increase the liability
of any director or officer of the Corporation with respect to any acts or omissions of such director or officer occurring prior to, such
amendment, repeal or modification.
Tenth:
To the fullest extent permitted by applicable law, the Corporation is authorized to provide indemnification of (and advancement of expenses
to) directors, officers and agents of the Corporation (and any other persons to which General Corporation Law permits the Corporation
to provide indemnification) through Bylaw provisions, agreements with such agents or other persons, vote of stockholders or disinterested
directors or otherwise, in excess of the indemnification and advancement otherwise permitted by Section 145 of the General Corporation
Law.
Any
amendment, repeal or modification of the foregoing provisions of this Article Tenth shall not (a) adversely affect any right or
protection of any director, officer or other agent of the Corporation existing at the time of such amendment, repeal or modification or
(b) increase the liability of any director of the Corporation with respect to any acts or omissions of such director, officer or agent
occurring prior to, such amendment, repeal or modification.
Eleventh:
The Corporation renounces, to the fullest extent permitted by law, any interest or expectancy of the Corporation in, or in being offered
an opportunity to participate in, any Excluded Opportunity. An “Excluded Opportunity” is any matter, transaction or
interest that is presented to, or acquired, created or developed by, or which otherwise comes into the possession of (i) any director
of the Corporation who is not an employee of the Corporation or any of its subsidiaries, or (ii) any holder of Preferred Stock or any
partner, member, director, stockholder, employee, affiliate or agent of any such holder, other than someone who is an employee of the
Corporation or any of its subsidiaries (collectively, the persons referred to in clauses (i) and (ii) are “Covered
Persons”), unless such matter, transaction or interest is presented to, or acquired, created or developed by, or otherwise comes
into the possession of, a Covered Person expressly and solely in such Covered Person’s capacity as a director of the Corporation
while such Covered Person is performing services in such capacity. Any repeal or modification of this Article Eleventh will only be prospective
and will not affect the rights under this Article Eleventh in effect at the time of the occurrence of any actions or omissions to act
giving rise to liability. Notwithstanding anything to the contrary contained elsewhere in this Amended and Restated Certificate of Incorporation,
the affirmative vote of the holders of a majority of the shares of Preferred Stock then outstanding, will be required to amend or repeal,
or to adopt any provisions inconsistent with this Article Eleventh.
Twelfth:
Unless the Corporation consents in writing to the selection of an alternative forum, the Court of Chancery in the State of Delaware shall
be the sole and exclusive forum for any stockholder (including a beneficial owner) to bring (i) any derivative action or proceeding
brought on behalf of the Corporation, (ii) any action asserting a claim of breach of fiduciary duty owed by any director, officer
or other employee of the Corporation to the Corporation or the Corporation’s stockholders, (iii) any action asserting a claim
against the Corporation, its directors, officers or employees arising pursuant to any provision of the Delaware General Corporation Law
or the Corporation’s certificate of incorporation or bylaws or (iv) any action asserting a claim against the Corporation, its
directors, officers or employees governed by the internal affairs doctrine, except for, as to each of (i) through (iv) above, any claim
as to which the Court of Chancery determines that there is an indispensable party not subject to the jurisdiction of the Court of Chancery
(and the indispensable party does not consent to the personal jurisdiction of the Court of Chancery within ten days following such determination),
which is vested in the exclusive jurisdiction of a court or forum other than the Court of Chancery, or for which the Court of Chancery
does not have subject matter jurisdiction. If any provision or provisions of this Article Twelfth shall be held to be invalid, illegal
or unenforceable as applied to any person or entity or circumstance for any reason whatsoever, then, to the fullest extent permitted by
law, the validity, legality and enforceability of such provisions in any other circumstance and of the remaining provisions of this Article
Twelfth (including, without limitation, each portion of any sentence of this Article Twelfth containing any such provision held to be
invalid, illegal or unenforceable that is not itself held to be invalid, illegal or unenforceable) and the application of such provision
to other persons or entities and circumstances shall not in any way be affected or impaired thereby.
Thirteenth:
For purposes of Section 500 of the California Corporations Code (to the extent applicable), in connection with any repurchase of shares
of Common Stock permitted under this Amended and Restated Certificate of Incorporation from employees, officers, directors or consultants
of the Corporation in connection with a termination of employment or services pursuant to agreements or arrangements approved by the Board
of Directors (in addition to any other consent required under this Amended and Restated Certificate of Incorporation), such repurchase
may be made without regard to any “preferential dividends arrears amount” or “preferential rights amount” (as
those terms are defined in Section 500 of the California Corporations Code). Accordingly, for purposes of making any calculation under
California Corporations Code Section 500 in connection with such repurchase, the amount of any “preferential dividends arrears amount”
or “preferential rights amount” (as those terms are defined therein) shall be deemed to be zero.
* * *
3.
That the foregoing amendment and restatement was approved by the holders of the requisite number of shares of this corporation
in accordance with Section 228 of the General Corporation Law.
4.
That this Certificate of Incorporation, which restates and integrates and further amends the provisions of this Corporation’s
Certificate of Incorporation, has been duly adopted in accordance with Sections 242 and 245 of the General Corporation Law.
[Signature Page Follows]
IN
WITNESS WHEREOF, this Amended and Restated Certificate of Incorporation has been executed by a duly authorized officer of this
corporation on this 20th day of May, 2026.
| |
By: |
/s/ Andrew Orth |
| |
|
Andrew Orth, President and CEO |
CERTIFICATE
OF AMENDMENT TO THE
AMENDED
AND RESTATED
CERTIFICATE
OF INCORPORATION
OF
CITY
THERAPEUTICS, INC.
(Pursuant to Section 242 of the
General Corporation Law of the State of Delaware)
City Therapeutics, Inc.,
a corporation organized and existing under and by virtue of the provisions of the General Corporation Law of the State of Delaware (the
“General Corporation Law”),
DOES
HEREBY CERTIFY:
FIRST:
That the name of this corporation is City Therapeutics, Inc., and that this corporation was originally incorporated pursuant to the
General Corporation Law on July 17, 2023.
SECOND:
Pursuant to Section 242 of the General Corporation Law, this Certificate of Amendment to the Amended and Restated Certificate of
Incorporation of this corporation (the “Certificate of Amendment”) amends the Amended and Restated Certificate of Incorporation
of this corporation (the “Restated Certificate of Incorporation”) as follows:
1. The
first paragraph of Article Fourth of the Restated Certificate of Incorporation be deleted and replaced in full with the following:
“The total number of shares of
all classes of stock which the Corporation shall have authority to issue is (i) 42,350,000 shares of Common Stock, $0.0001 par value
per share (“Common Stock”) and (ii) 26,266,656 shares of Preferred Stock, $0.0001 par value per share (“Preferred
Stock”).”
THIRD:
That the Board of Directors of the Corporation duly approved the Certificate of Amendment in accordance with the provisions of Sections
141 and 242 of the General Corporation Law.
FOURTH:
The holders of the requisite number of shares of the Corporation approved and adopted the Certificate of Amendment in accordance with
Sections 228 and 242 of the General Corporation Law.
FIFTH:
Other than as set forth in this Certificate of Amendment, the Restated Certificate of Incorporation, as amended, shall remain in full
force and effect, without modification, amendment or change.
* * *
IN
WITNESS WHEREOF, this Certificate of Amendment to the Amended and Restated Certificate of Incorporation has been executed by
a duly authorized officer of this corporation on June 4, 2026.
| |
/s/ Andrew Orth |
| |
Andrew Orth, President and CEO |
CERTIFICATE OF AMENDMENT TO THE
AMENDED AND
RESTATED
CERTIFICATE
OF INCORPORATION
OF
CITY THERAPEUTICS,
INC.
(Pursuant to Section 242 of
the
General Corporation Law of
the State of Delaware)
City Therapeutics, Inc. (the
“Corporation”), a corporation organized and existing under and by virtue of the provisions of the General Corporation
Law of the State of Delaware (the “General Corporation Law”),
DOES HEREBY CERTIFY:
1. That
the name of the Corporation is City Therapeutics, Inc., and that the Corporation was originally incorporated pursuant to the General
Corporation Law on July 17, 2023 under the name City Therapeutics, Inc. (the “Original Certificate”). The Original
Certificate was amended and restated by that certain Amended and Restated Certificate of Incorporation dated May 20, 2026, and further
amended by that certain Certificate of Amendment to the Amended and Restated Certificate of Incorporation dated June 4, 2026 (as
amended, the “Amended and Restated Certificate of Incorporation”).
2. That
the Board of Directors of the Corporation duly adopted resolutions proposing to amend the Amended and Restated Certificate of Incorporation,
declaring said amendments to be advisable and in the best interests of the Corporation and its stockholders, and authorizing the appropriate
officers of the Corporation to solicit the consent of the stockholders therefor, which resolutions setting forth the proposed amendments
are as follows:
RESOLVED, that the
following is hereby inserted into Article FOURTH immediately before the first sentence therein:
“Effective
upon the filing of this Certificate of Amendment to the Amended and Restated Certificate of Incorporation with the Secretary of State
of the State of Delaware (the “Effective Time”), every one (1) share of Common Stock then issued and outstanding or
held in the treasury of the Corporation immediately prior to the Effective Time shall automatically be subdivided into 1.0983 shares of
Common Stock, without any further action by the holders of such shares (the “Forward Stock Split”). The Forward Stock
Split will be effected on a certificate-by-certificate basis, and any fractional shares resulting from such combination shall be rounded
down to the nearest whole share on a certificate-by-certificate basis. No fractional shares shall be issued in connection with the Forward
Stock Split. In lieu of any fractional shares to which a holder would otherwise be entitled, the Corporation shall pay cash equal to such
fraction multiplied by the fair market value of a share of Common Stock as determined in good faith by the Corporation’s Board of
Directors. The Forward Stock Split shall occur automatically without any further action by the holders of the shares of Common Stock and
Preferred Stock affected thereby. All rights, preferences and privileges of the Common Stock and the Preferred Stock shall be appropriately
adjusted to reflect the Forward Stock Split in accordance with this Amended and Restated Certificate of Incorporation.”
RESOLVED, that Section
4.2 of Part B of Article FOURTH of the Amended and Restated Certificate of Incorporation be amended and restated in its entirety to read
as follows:
“4.2 Fractional
Shares. No fractional shares of Common Stock shall be issued upon conversion of the Preferred Stock. In lieu of any fractional shares
to which the holder would otherwise be entitled, the Corporation shall pay cash equal to such fraction multiplied by the fair market value
of a share of Common Stock as determined in good faith by the Board of Directors. Whether or not fractional shares would be issuable upon
such conversion shall be determined on the basis of (i) the total number of shares of Preferred Stock formerly represented prior to such
conversion by a stock certificate (or book-entry position) that the holder surrenders for conversion and (ii) the number of shares of
Common Stock after such conversion into which such shares of Preferred Stock formerly represented by such stock certificate (or book-entry
position) converted.”
RESOLVED, that the
final sentence of Section 4.3.1 of Part B of Article FOURTH of the Amended and Restated Certificate of Incorporation be amended and restated
in its entirety to read as follows:
“The Corporation
shall, as soon as practicable after the Conversion Time (i) issue and deliver to such holder of Preferred Stock, or to his, her or its
nominees, a certificate or certificates (or book-entry accounts) for the number of full shares of Common Stock issuable upon such conversion
in accordance with the provisions hereof and a certificate for the number (if any) of the shares of Preferred Stock represented by the
surrendered certificate that were not converted into Common Stock, (ii) pay in cash such amount as provided in Section 4.2 in lieu of
any fraction of a share of Common Stock otherwise issuable upon such conversion and (iii) pay all declared but unpaid dividends on the
shares of Preferred Stock converted.”
RESOLVED, that the
first sentence of Section 4.3.3 of Part B of Article FOURTH of the Amended and Restated Certificate of Incorporation be amended and restated
in its entirety to read as follows:
“All shares
of Preferred Stock which shall have been surrendered for conversion as herein provided shall no longer be deemed to be outstanding and
all rights with respect to such shares shall immediately cease and terminate at the Conversion Time, except only the right of the holders
thereof to receive shares of Common Stock in exchange therefor, to receive payment in lieu of any fraction of a share otherwise issuable
upon such conversion as provided in Section 4.2 and to receive payment of any dividends declared but unpaid thereon.”
RESOLVED, that the
sixth sentence of Section 5.2 of Part B of Article FOURTH of the Amended and Restated Certificate of Incorporation be amended and restated
in its entirety to read as follows:
“As soon as
practicable after the Mandatory Conversion Time and, if applicable, the surrender of any certificate or certificates (or lost certificate
affidavit and agreement) for Preferred Stock, the Corporation shall (a) issue and deliver to such holder, or to his, her or its nominees,
a certificate or certificates (or book-entry accounts) for the number of full shares of Common Stock issuable on such conversion in accordance
with the provisions hereof or issue and deliver to such holder, or to his, her or its nominees, a notice of issuance of uncertificated
shares and may, upon written request, issue and deliver a certificate for the number of full shares of Common Stock issuable upon such
conversion in accordance with the provisions hereof, (b) pay in cash such amount as provided in Section 4.2 in lieu of any fraction of
a share of Common Stock otherwise issuable upon such conversion, and (c) pay any declared but unpaid dividends on the shares of Preferred
Stock converted.”
3. That
the foregoing amendments were approved by the holders of the requisite number of shares of the Corporation in accordance with Section
228 of the General Corporation Law.
4. That said amendments have been duly adopted in accordance with Section 242 of the General Corporation Law.
5. All
other provisions of the Amended and Restated Certificate of Incorporation shall remain in full force and effect.
[Signature Page to Follow]
IN
WITNESS WHEREOF, this Certificate of Amendment to the Amended and Restated Certificate of Incorporation has been executed by
a duly authorized officer of the Corporation on October 8, 2026.
| |
/s/
Andrew Orth |
| |
Name: Andrew Orth |
| |
Title: President and Chief Executive Officer |
Exhibit 3.2
AMENDED AND RESTATED
CERTIFICATE OF INCORPORATION
OF
CITY THERAPEUTICS, INC.
City Therapeutics, Inc., a corporation organized
and existing under the laws of the State of Delaware (the “Corporation”), hereby certifies as follows:
1. The
name of the Corporation is City Therapeutics, Inc. The date of the filing of its original Certificate of Incorporation with the Secretary
of State of the State of Delaware was July 17, 2023 (the “Original Certificate”). The name under which the Corporation
filed the Original Certificate was City Therapeutics, Inc.
2. This
Amended and Restated Certificate of Incorporation (the “Certificate”) amends, restates and integrates the provisions
of the Amended and Restated Certificate of Incorporation that was filed with the Secretary of State of the State of Delaware on May 20,
2026, as further amended by that certain Certificate of Amendment to the Amended and Restated Certificate of Incorporation that was filed
with the Secretary of State of the State of Delaware on June 4, 2026 (as amended, the “Amended and Restated Certificate”),
and was duly adopted in accordance with the provisions of Sections 228, 242 and 245 of the General Corporation Law of the State of Delaware
(the “DGCL”).
3. The
text of the Amended and Restated Certificate is hereby amended, restated and integrated in its entirety to provide as follows.
ARTICLE I
The name of the Corporation is City Therapeutics, Inc.
ARTICLE II
The address of the Corporation’s registered
office in the State of Delaware is 3500 South DuPont Highway in the City of Dover, County of Kent, Delaware 19901. The name of its registered
agent at such address is Incorporating Services, Ltd.
ARTICLE III
The purpose of the Corporation is to engage in
any lawful act or activity for which corporations may be organized under the DGCL.
ARTICLE IV
CAPITAL STOCK
The total number of shares of capital stock which
the Corporation shall have authority to issue is five hundred and ten million (510,000,000), of which (i) five hundred million (500,000,000)
shares shall be a class designated as common stock, par value $0.0001 per share (the “Common Stock”), and (ii) ten
million (10,000,000) shares shall be a class designated as undesignated preferred stock, par value $0.0001 per share (the “Preferred
Stock”).
Except as otherwise provided in any certificate
of designation of any series of Preferred Stock, the number of authorized shares of the class of Common Stock or Preferred Stock may be
increased or decreased (but not below the number of shares of such class then outstanding) by the affirmative vote of the holders of a
majority in voting power of the outstanding shares of capital stock of the Corporation entitled to vote thereon irrespective of the provisions
of Section 242(b)(2) of the DGCL, and no vote of the holders of any of the Common Stock or the Preferred Stock voting separately
as a class shall be required therefor. For the avoidance of doubt, the elimination and reduction of the voting requirements of Section 242
of the DGCL, as permitted by Section 242(d) of the DGCL, shall apply to any amendments to the Amended and Restated Certificate
of Incorporation (the “Certificate”).
The powers, preferences and rights of, and the
qualifications, limitations and restrictions upon, each class or series of stock shall be determined in accordance with, or as set forth
below in, this Article IV.
A. COMMON STOCK
Subject to all the rights, powers and preferences
of the Preferred Stock and except as provided by law or in this Certificate (including any certificate of designation of any series of
Preferred Stock):
(a) the
holders of the Common Stock shall have the exclusive right to vote for the election of directors of the Corporation (the “Directors”)
and on all other matters requiring stockholder action, each outstanding share entitling the holder thereof to one vote on each matter
properly submitted to the stockholders of the Corporation for their vote; provided, however, that, except as otherwise required
by law, holders of Common Stock, as such, shall not be entitled to vote on any amendment to this Certificate (including any amendment
to a certificate of designation of any series of Preferred Stock) that alters or changes the powers, preferences, rights or other terms
of one or more outstanding series of Preferred Stock if the holders of such affected series of Preferred Stock are entitled to vote, either
separately or together with the holders of one or more other such series, on such amendment pursuant to this Certificate (including any
certificate of designation of any series of Preferred Stock) or pursuant to the DGCL;
(b) dividends
may be declared and paid or set apart for payment upon the shares of Common Stock out of any assets or funds of the Corporation legally
available for the payment of dividends, but only when, as and if declared by the Board of Directors of the Corporation (the “Board
of Directors”) or any authorized committee thereof; and
(c) upon
the voluntary or involuntary liquidation, dissolution or winding up of the Corporation, the net assets of the Corporation shall be distributed
pro rata to the holders of the Common Stock.
B. PREFERRED STOCK
The Board of Directors or any authorized committee
thereof is expressly authorized to provide by resolution or resolutions for, out of the unissued shares of Preferred Stock, the issuance
of the shares of Preferred Stock in one or more series of such stock, and by filing a certificate of designation pursuant to applicable
law of the State of Delaware, to establish or change from time to time the number of shares of each such series, and to fix the designations,
powers, including voting powers, full or limited, or no voting powers, preferences and the relative, participating, optional or other
special rights of the shares of each series and any qualifications, limitations and restrictions thereof, all to the fullest extent now
or hereafter permitted by the DGCL. The powers, preferences and relative, participating, optional and other special rights of each such
series of Preferred Stock, and the qualifications, limitations or restrictions thereof, if any, may differ from those of any and all other
series at any time outstanding. Without limiting the generality of the foregoing, the resolution or resolutions providing for the issuance
of any series of Preferred Stock may provide that such series shall be superior or rank equally or be junior to any other series of Preferred
Stock to the extent permitted by law.
ARTICLE V
STOCKHOLDER ACTION
1. Action
without Meeting. Subject to the rights, if any, of the holders of shares of any series of Preferred Stock, any action required or
permitted to be taken by the stockholders of the Corporation at any annual or special meeting of stockholders of the Corporation must
be effected at a duly called annual or special meeting of stockholders and may not be taken or effected by a consent of stockholders in
lieu thereof.
2. Special
Meetings. Except as otherwise required by statute and subject to the rights, if any, of the holders of shares of any series of Preferred
Stock, special meetings of the stockholders of the Corporation may be called only by the Board of Directors, and special meetings of stockholders
may not be called by any other person or persons. Only those matters set forth in the notice of the special meeting may be considered
or acted upon at a special meeting of stockholders of the Corporation.
ARTICLE VI
DIRECTORS
1. General.
The business and affairs of the Corporation shall be managed by or under the direction of the Board of Directors except as otherwise provided
herein or required by law.
2. Number
of Directors; Term of Office. Except as otherwise provided for or fixed pursuant to the provisions of Article IV (including any
certificate of designation with respect to any series of Preferred Stock) and this Article VII relating to the rights of the holders
of any series of Preferred Stock to elect additional Directors, the number of Directors shall be fixed solely and exclusively by resolution
duly adopted from time to time by the Board of Directors. The Directors, other than those who may be elected by the holders of any series
of Preferred Stock, shall be classified, with respect to the term for which they severally hold office, into three classes. The term of
office of the initial Class I Directors shall expire at the first regularly-scheduled annual meeting of stockholders following the
closing of the Corporation’s sale of a class of its capital stock to the public pursuant to a registration statement on Form S-1
under the Securities Act (the “IPO Time”). The term of office of the initial Class II Directors shall expire at
the second annual meeting of stockholders following the IPO Time. The term of office of the initial Class III Directors shall expire
at the third annual meeting of stockholders following the IPO Time. The Board of Directors is authorized to assign members of the Board
of Directors already in office to such classes at the time the classification of the Board of Directors becomes effective. At each annual
meeting of stockholders, Directors elected to succeed those Directors whose terms expire shall be elected for a term of office to expire
at the third succeeding annual meeting of stockholders after their election. Notwithstanding the foregoing, the Directors elected to each
class shall hold office until their successors are duly elected and qualified or until their earlier resignation, death, disqualification
or removal. No decrease in the number of Directors shall shorten the term of any incumbent Director. There shall be no cumulative voting
in the election of Directors. Election of Directors need not be by written ballot unless the Bylaws of the Corporation so provide.
Notwithstanding the foregoing, whenever, pursuant
to the provisions of Article IV of this Certificate, the holders of any one or more series of Preferred Stock shall have the right,
voting separately as a series or together with holders of other such series, to elect additional Directors, the election, term of office,
filling of vacancies and other features of such directorships shall be governed by the terms of this Certificate, including any certificate
of designation applicable to such series of Preferred Stock. During any period when the holders of any series of Preferred Stock, voting
separately as a series or together with one or more series, have the right to elect additional Directors, then upon commencement and for
the duration of the period during which such right continues: (i) the then otherwise total authorized number of Directors shall automatically
be increased by such specified number of Directors, and the holders of such Preferred Stock shall be entitled to elect the additional
Directors so provided for or fixed pursuant to said provisions, and (ii) each such additional Director shall serve until such Director’s
successor shall have been duly elected and qualified, or until such Director’s right to hold such office terminates pursuant to
said provisions, whichever occurs earlier, subject to such Director’s earlier death, resignation, retirement, disqualification or
removal. Notwithstanding any other provision of this Certificate of Incorporation, except as otherwise provided by the Board of Directors
in the resolution or resolutions establishing such series, whenever the holders of any series of Preferred Stock having such right to
elect additional Directors are divested of such right pursuant to the provisions of such stock, the terms of office of all such additional
Directors elected by the holders of such stock, or elected to fill any vacancies resulting from the death, resignation, disqualification
or removal of such additional Directors, shall forthwith terminate (in which case each such Director shall thereupon cease to be qualified
as, and shall cease to be, a Director) and the total authorized number of Directors shall automatically be reduced accordingly.
3. Vacancies
and Newly Created Directorships. Subject to the rights, if any, of the holders of any series of Preferred Stock to elect Directors
and to fill vacancies in the Board of Directors relating thereto, any and all vacancies and newly created directorships in the Board of
Directors, however occurring, including, without limitation, by reason of an increase in the size of the Board of Directors, or the death,
resignation, disqualification or removal of a Director, shall be filled solely and exclusively by the affirmative vote of a majority of
the remaining Directors then in office, even if less than a quorum of the Board of Directors, or by a sole remaining Director, and not
by the stockholders. Any Director appointed in accordance with the preceding sentence shall hold office for the remainder of the full
term of the class of Directors in which the new directorship was created or the vacancy occurred and until such Director’s successor
shall have been duly elected and qualified or until such Director’s earlier resignation, disqualification, death or removal. Subject
to the rights, if any, of the holders of any series of Preferred Stock to elect Directors, when the number of Directors is increased or
decreased, the Board of Directors shall, subject to Article VI.3 hereof, determine the class or classes to which the increased or
decreased number of Directors shall be apportioned. In the event of a vacancy in the Board of Directors, the remaining Directors, except
as otherwise provided by law, shall exercise the powers of the full Board of Directors until the vacancy is filled.
4. Removal.
Subject to the rights, if any, of any series of Preferred Stock to elect Directors and to remove any Director whom the holders of any
such series have the right to elect, any Director may be removed from office (i) only for cause and (ii) only by the affirmative
vote of the holders not less than two-thirds (2/3) of the voting power of the outstanding shares of capital stock then entitled to vote
at an election of Directors.
ARTICLE VII
LIMITATION OF LIABILITY
1. Directors.
To the fullest extent permitted by the DGCL, as the same exists or may hereafter be amended from time to time, a Director of the Corporation
shall not be personally liable to the Corporation or its stockholders for monetary damages for breach of such Director’s fiduciary
duty as a Director, except for liability (a) for any breach of the Director’s duty of loyalty to the Corporation or its stockholders,
(b) for acts or omissions not in good faith or which involve intentional misconduct or a knowing violation of law, (c) under
Section 174 of the DGCL or (d) for any transaction from which the Director derived an improper personal benefit. If the DGCL
is amended after the effective date of this Certificate to authorize corporate action further eliminating or limiting the personal liability
of Directors, then the liability of a Director of the Corporation shall be eliminated or limited to the fullest extent permitted by the
DGCL, as so amended.
2. Officers.
To the fullest extent permitted by the DGCL, as the same exists or may thereafter be amended from time to time, an Officer (as defined
below) of the Corporation shall not be personally liable to the Corporation or its stockholders for monetary damages for breach of such
Officer’s fiduciary duty as an officer of the Corporation, except for liability (a) for any breach of the Officer’s duty
of loyalty to the Corporation or its stockholders, (b) for acts or omissions not in good faith or which involve intentional misconduct
or a knowing violation of law, (c) for any transaction from which the Officer derived an improper personal benefit, or (d) arising
from any claim brought by or in the right of the Corporation. If the DGCL is amended after the effective date of this Certificate to authorize
corporate action further eliminating or limiting the personal liability of Officers, then the liability of an Officer of the Corporation
shall be eliminated or limited to the fullest extent permitted by the DGCL, as so amended. For purposes of this Article VII, “Officer”
shall mean an individual who has been duly appointed as an officer of the Corporation and who, at the time of an act or omission as to
which liability is asserted, is deemed to have consented to service by the delivery of process to the registered agent of the Corporation
as contemplated by 10 Del. C. § 3114(b).
3. Amendment
or Modification. Any amendment, repeal or modification of this Article VII or any amendment to the DGCL, shall not adversely
affect any right or protection existing at the time of such amendment, repeal or modification with respect to any acts or omissions occurring
before such amendment, repeal or modification of a person serving as a Director or Officer, as applicable, at the time of such amendment,
repeal or modification.
ARTICLE VIII
AMENDMENT OF BYLAWS
1. Amendment
by Directors. Except as otherwise provided by law, the Bylaws of the Corporation may be adopted, amended or repealed by the Board
of Directors.
2. Amendment
by Stockholders. Except as otherwise provided therein, the Bylaws of the Corporation may be amended or repealed by the stockholders
by the affirmative vote of the holders of at least two-thirds (2/3) of the voting power of the outstanding shares of capital stock entitled
to vote on such amendment or repeal, voting together as a single class; provided, however, that if the Board of Directors recommends that
stockholders approve such amendment or repeal, such amendment or repeal shall only require the affirmative vote of the holders of a majority
of the voting power of the outstanding shares of capital stock entitled to vote on such amendment or repeal, voting together as a single
class.
ARTICLE IX
AMENDMENT OF CERTIFICATE OF INCORPORATION
The Corporation reserves the right to amend or
repeal this Certificate in the manner now or hereafter prescribed by statute and this Certificate, and all rights conferred upon stockholders
herein are granted subject to this reservation. For the avoidance of doubt, the provisions of Sections 242(d)(1) and (d)(2) of
the DGCL shall apply to the Corporation.
[Signature Page Follows]
THIS AMENDED AND RESTATED
CERTIFICATE OF INCORPORATION is executed as of this [·]th day of [·], 2026.
| |
CITY THERAPEUTICS, INC. |
| |
|
|
| |
By: |
|
| |
Name: Andrew Orth |
| |
Title: President and Chief Executive Officer |
[Signature Page to Amended and Restated Certificate
of Incorporation]
Exhibit 3.4
AMENDED AND RESTATED
BYLAWS
OF
CITY THERAPEUTICS, INC.
(the “Corporation”)
Article I
Stockholders
Section 1. Annual
Meeting. The annual meeting of stockholders (any such meeting being referred to in these Bylaws as an “Annual Meeting”)
shall be held at the hour, date and place within or without the United States that is fixed by or in the manner determined by the Board
of Directors and stated in the notice of the meeting, which time, date and place may subsequently be changed at any time, before or after
the notice for such meeting has been sent to the stockholders, by vote of the Board of Directors. The Board of Directors may, in its
sole discretion, determine that a meeting of stockholders shall not be held at any place, but may instead be held solely by means of
remote communication as authorized by Section 211(a)(2) of the General Corporation Law of the State of Delaware (the “DGCL”).
In the absence of any such designation or determination, stockholders’ meetings shall be held at the Corporation’s principal
executive office. If no Annual Meeting has been held for a period of thirteen (13) months after the Corporation’s last Annual Meeting,
a special meeting in lieu thereof may be held, and such special meeting shall have, for the purposes of these Bylaws or otherwise, all
the force and effect of an Annual Meeting. Any and all references hereafter in these Bylaws to an Annual Meeting or Annual Meetings also
shall be deemed to refer to any special meeting(s) in lieu thereof.
Section 2. Notice
of Stockholder Business and Nominations.
(a) Annual
Meetings of Stockholders.
(1) Nominations
of persons for election to the Board of Directors of the Corporation (the “Board of Directors”) and the proposal of
other business to be considered by the stockholders may be brought before an Annual Meeting (i) by or at the direction of the Board
of Directors or (ii) by any stockholder of the Corporation who was a stockholder of record at the time of giving of notice of the
Annual Meeting provided for in this Bylaw, who is entitled to vote at the meeting, who is present (in person or by proxy) at the meeting
and who complies with the notice procedures set forth in this Bylaw as to such nomination or business. For the avoidance of doubt, the
foregoing clause (ii) shall be the exclusive means for a stockholder to bring nominations or business properly before an Annual
Meeting (other than matters properly brought under Rule 14a-8 (or any successor rule) under the Securities Exchange Act of 1934,
as amended (the “Exchange Act”)), and such stockholder must comply with the notice and other procedures set forth
in Article I, Section 2(a)(2), (3) and (4) of this Bylaw to bring such nominations or business properly before an
Annual Meeting. In addition to the other requirements set forth in this Bylaw, for any proposal of business to be considered at an Annual
Meeting, it must be a proper subject for action by stockholders of the Corporation under Delaware law.
(2) For
nominations or other business to be properly brought before an Annual Meeting by a stockholder pursuant to clause (ii) of Article I,
Section 2(a)(1) of this Bylaw, the stockholder must (i) have given Timely Notice (as defined below) thereof in writing
to the Secretary of the Corporation, (ii) have provided any updates or supplements to such notice at the times and in the forms
required by this Bylaw and (iii) together with the beneficial owner(s), if any, on whose behalf the nomination or business proposal
is made, have acted in accordance with the representations set forth in the Solicitation Statement (as defined below) required by this
Bylaw. To be timely, a stockholder’s written notice must be received by the Secretary at the principal executive offices of the
Corporation not later than 5:00 p.m. Eastern time on the ninetieth (90th) day nor earlier than 5:00 p.m. Eastern Time on the
one hundred twentieth (120th) day prior to the one-year anniversary of the preceding year’s Annual Meeting; provided, however,
that in the event the Annual Meeting is first convened more than thirty (30) days before or more than sixty (60) days after such anniversary
date, or if no Annual Meeting was held in the preceding year, notice by the stockholder to be timely must be received by the Secretary
of the Corporation not later than 5:00 p.m. Eastern time on the later of the ninetieth (90th) day prior to the scheduled date of
such Annual Meeting or the tenth (10th) day following the day on which public announcement of the date of such meeting is first made
(such notice within such time periods shall be referred to as “Timely Notice”). Notwithstanding anything to the contrary
provided herein, for the first Annual Meeting following the initial public offering of common stock of the Corporation, a stockholder’s
notice shall be timely if received by the Secretary at the principal executive offices of the Corporation not later than 5:00 p.m. Eastern
time on the later of the ninetieth (90th) day prior to the scheduled date of such Annual Meeting or the tenth (10th) day following the
day on which public announcement of the date of such Annual Meeting is first made or sent by the Corporation. Such stockholder’s
Timely Notice shall set forth or include:
(A) as
to each person whom the stockholder proposes to nominate for election or reelection as a director, (i) the name, age, business address
and residence address of the nominee, (ii) the principal occupation or employment of the nominee, (iii) the class and number
of shares of capital stock of the Corporation that are held of record or are beneficially owned by the nominee or its Affiliates or Associates
(each as defined below) and any Synthetic Equity Interest (as defined below) held or beneficially owned by the nominee or its Affiliates
or Associates, (iv) a description of all agreements, arrangements or understandings between or among the stockholder and each nominee
and any other person or persons (naming such person or persons) pursuant to which the nominations are to be made by the stockholder or
concerning the nominee’s potential service on the Board of Directors, (v) a questionnaire with respect to the background and
qualifications of the nominee completed by the nominee in the form provided by the Corporation (which questionnaire shall be provided
by the Secretary upon written request of any stockholder of record identified by name within five (5) business days of such written
request), (vi) a representation and agreement in the form provided by the Corporation (which form shall be provided by the Secretary
upon written request of any stockholder of record identified by name within five (5) business days of such written request) that:
(a) such proposed nominee is not and will not become party to any agreement, arrangement or understanding with any person or entity
as to how such proposed nominee, if elected as a director of the Corporation, will act or vote on any issue or question (a “Voting
Commitment”) that has not been disclosed to the Corporation in the questionnaire described in clause (v) herein; (b) such
proposed nominee is not and will not become a party to any agreement, arrangement or understanding with any person or entity other than
the Corporation with respect to any direct or indirect compensation, reimbursement or indemnification in connection with service or action
as a director that has not been disclosed to the Corporation in the questionnaire described in clause (v) herein; (c) such
proposed nominee would, if elected as a director, comply with all applicable rules and regulations of the exchanges upon which shares
of the Corporation’s capital stock trade, each of the Corporation’s corporate governance, ethics, conflict of interest, confidentiality,
stock ownership and trading policies and guidelines applicable generally to the Corporation’s directors and, if elected as a director
of the Corporation, such person currently would be in compliance with any such policies and guidelines that have been publicly disclosed;
(d) such proposed nominee intends to serve as a director for the full term for which he or she is to stand for election; and (e) such
proposed nominee will promptly provide to the Corporation such other information as it may reasonably request to determine the eligibility
of such proposed nominee to serve on any committee or sub-committee of the Board of Directors under any applicable stock exchange listing
requirements or applicable law, or that the Board of Directors reasonably determines could be material to a reasonable stockholder’s
understanding of the background, qualifications, experience, independence, or lack thereof, of such proposed nominee; and (vii) any
other information relating to such proposed nominee that is required to be disclosed in solicitations of proxies for election of directors
in an election contest, or is otherwise required, in each case pursuant to Regulation 14A under the Exchange Act (including, without
limitation, such person’s written consent to being named in the proxy statement as a nominee and to serving as a director if elected);
(B) as
to any other business that the stockholder proposes to bring before the meeting: a brief description of the business desired to be brought
before the meeting, the reasons for conducting such business at the meeting, the text, if any, of any resolutions or Bylaw amendment
proposed for adoption, and any material interest in such business of each Proposing Person (as defined below);
(C) (i) the
name and address of the stockholder giving the notice, as they appear on the Corporation’s books, and the names and addresses of
the other Proposing Persons (if any) and (ii) as to each Proposing Person, the following information: (a) the class or series
and number of all shares of capital stock of the Corporation that are, directly or indirectly, owned beneficially or of record by such
Proposing Person or any of its Affiliates or Associates, including any shares of any class or series of capital stock of the Corporation
as to which such Proposing Person or any of its Affiliates or Associates has a right to acquire beneficial ownership at any time in the
future (whether or not such right is exercisable immediately or only after the passage of time or upon the satisfaction of any conditions
or both) pursuant to any agreement, arrangement or understanding (whether or not in writing), (b) all Synthetic Equity Interests
(as defined below) in which such Proposing Person or any of its Affiliates or Associates, directly or indirectly, holds an interest including
a description of the material terms of each such Synthetic Equity Interest, including, without limitation, identification of the counterparty
to each such Synthetic Equity Interest and disclosure, for each such Synthetic Equity Interest, as to (1) whether or not such Synthetic
Equity Interest conveys any voting rights, directly or indirectly, in such shares to such Proposing Person or any of its Affiliates or
Associates and (2) whether or not such Synthetic Equity Interest is required to be, or is capable of being, settled through delivery
of such shares, (c) any proxy (other than a revocable proxy given in response to a public proxy solicitation made pursuant to, and
in accordance with, the Exchange Act), agreement, arrangement, understanding or relationship pursuant to which such Proposing Person
or any of its Affiliates or Associates has or shares a right to, directly or indirectly, vote any shares of any class or series of capital
stock of the Corporation, (d) any rights to dividends or other distributions on the shares of any class or series of capital stock
of the Corporation, directly or indirectly, owned beneficially by such Proposing Person or any of its Affiliates or Associates that are
separated or separable from the underlying shares of the Corporation, (e) if such Proposing Person is not a natural person, the
identity of the natural person or persons responsible for making voting and investment decisions (including director nominations and
any other business that the stockholder proposes to bring before a meeting) on behalf of the Proposing Person (irrespective of whether
such person or persons have “beneficial ownership” for purposes of Rule 13d-3 of the Exchange Act of any securities
owned of record or beneficially by the Proposing Person) (such person or persons, the “Responsible Person”), (f) any
pending or threatened litigation in which such Proposing Person or any of its Affiliates or Associates or any Responsible Person is a
party involving the Corporation or any of its officers or directors, or any Affiliate of the Corporation, and (g) any other information
relating to such Proposing Person or any of its Affiliates or Associates that would be required to be disclosed in a proxy statement
or other filing required to be made in connection with solicitations of proxies or consents by such Proposing Person in support of the
business proposed to be brought before the meeting pursuant to Section 14(a) of the Exchange Act (the disclosures to be made
pursuant to the foregoing clauses (a) through (g) are referred to, collectively, as “Material Ownership Interests”);
provided, however, that the Material Ownership Interests shall not include any such disclosures with respect to the ordinary course business
activities of any broker, dealer, commercial bank, trust company or other nominee who is a Proposing Person solely as a result of being
the stockholder of record directed to prepare and submit the notice required by these Bylaws on behalf of a beneficial owner;
(D) (i) a
description of all agreements, arrangements or understandings to which any Proposing Person or any of its Affiliates or Associates is
a party (whether the counterparty or counterparties are a Proposing Person or any Affiliate or Associate thereof, on the one hand, or
one or more other third parties, on the other hand, (including any proposed nominee(s)) (a) pertaining to the nomination(s) or
other business proposed to be brought before the meeting of stockholders or (b) entered into for the purpose of acquiring, holding,
disposing or voting of any shares of any class or series of capital stock of the Corporation (which description shall identify the name
of each other person who is party to such an agreement, arrangement or understanding) and (ii) identification of the names and addresses
of other stockholders (including beneficial owners) known by any of the Proposing Persons to be providing financial support or meaningful
assistance in furtherance of the nomination(s) or other business proposed to be brought before the meeting of stockholders and,
to the extent known, the class and number of all shares of the Corporation’s capital stock owned beneficially or of record by such
other stockholder(s) or other beneficial owner(s); and
(E) a
statement (i) that the stockholder is a holder of record of capital stock of the Corporation entitled to vote at such meeting, a
representation that such stockholder intends to appear in person or by proxy at the meeting to propose such business or nominees and
an acknowledgement that, if such stockholder (or a qualified representative of such stockholder) does not appear to present such business
or proposed nominees, as applicable, at such meeting, the Corporation need not present such business or proposed nominees for a vote
at such meeting, notwithstanding that proxies in respect of such vote may have been received by the Corporation, (ii) whether or
not the stockholder giving the notice and/or the other Proposing Person(s), if any, (a) will deliver a proxy statement and form
of proxy to holders of, in the case of a business proposal, at least the percentage of voting power of all of the shares of capital stock
of the Corporation required under applicable law to approve the proposal or, in the case of a nomination or nominations, at least 67
percent of the voting power of all of the shares of capital stock of the Corporation entitled to vote on the election of directors or
(b) otherwise solicit proxies or votes from stockholders in support of such proposal or nomination, as applicable, (iii) providing
a representation as to whether or not such Proposing Person intends to solicit proxies in support of director nominees other than the
Corporation’s director nominees in accordance with Rule 14a-19 promulgated under the Exchange Act and (iv) that the stockholder
will provide any other information relating to such item of business that would be required to be disclosed in a proxy statement or other
filing required to be made in connection with solicitations of proxies in support of the business proposed to be brought before the meeting
pursuant to Section 14(a) of the Exchange Act (such statement, the “Solicitation Statement”).
For purposes of this Article I,
the term “Proposing Person” shall mean the following persons: (i) the stockholder of record providing the notice of
nominations or business proposed to be brought before a stockholders’ meeting and (ii) the beneficial owner(s), if different,
on whose behalf the nominations or business proposed to be brought before a stockholders’ meeting is made. For purposes of this
Section 2, each of the terms “Affiliates” and “Associates” shall have the meaning attributed to such term
in Rule 12b-2 under the Exchange Act. For purposes of this Section 2, the term “Synthetic Equity Interest” shall
mean any transaction, agreement or arrangement (or series of transactions, agreements or arrangements), including, without limitation,
any derivative, swap, hedge, repurchase or so-called “stock borrowing” or securities lending agreement or arrangement, the
purpose or effect of which is to, directly or indirectly: (a) give a person or entity economic benefit and/or risk similar to ownership
of shares of any class or series of capital stock of the Corporation, in whole or in part, including due to the fact that such transaction,
agreement or arrangement provides, directly or indirectly, the opportunity to profit, or share in any profit, or avoid a loss from any
increase or decrease in the value of any shares of any class or series of capital stock of the Corporation, (b) mitigate loss to,
reduce the economic risk of, or manage the risk of share price changes for, any person or entity with respect to any shares of any class
or series of capital stock of the Corporation, or (c) increase or decrease the voting power of any person or entity with respect
to any shares of any class or series of capital stock of the Corporation.
(3) A
stockholder providing Timely Notice of nominations or business proposed to be brought before an Annual Meeting shall further update and
supplement such notice, if necessary, so that the information (including, without limitation, the Material Ownership Interests information)
provided or required to be provided in such notice pursuant to this Bylaw shall be true and correct as of the record date for the meeting
and as of the date that is ten (10) business days prior to such Annual Meeting, and such update and supplement shall be received
by the Secretary at the principal executive offices of the Corporation not later than 5:00 p.m. Eastern time on the fifth (5th)
business day after the record date for the Annual Meeting (in the case of the update and supplement required to be made as of the record
date), and not later than 5:00 p.m. Eastern time on the eighth (8th) business day prior to the date of the Annual Meeting (in the
case of the update and supplement required to be made as of ten (10) business days prior to the meeting). For the avoidance of doubt,
the obligation to update as set forth in this Section 2(a)(3) shall not limit the Corporation’s rights with respect to
any deficiencies in any notice provided by a stockholder, extend any applicable deadlines hereunder, or enable or be deemed to permit
a stockholder who has previously submitted notice hereunder to amend or update any proposal or nomination or to submit any new proposal,
including by changing or adding nominees, matters, business and/or resolutions proposed to be brought before a meeting of the stockholders.
Notwithstanding the foregoing, if a Proposing Person no longer plans to solicit proxies in accordance with its representation pursuant
to Article I, Section 2(a)(2)(E), such Proposing Person shall inform the Corporation of this change by delivering a written
notice to the Secretary at the principal executive offices of the Corporation no later than two (2) business days after making the
determination not to proceed with a solicitation of proxies. A Proposing Person shall also update its notice so that the information
required by Article I, Section 2(a)(2)(C) is current through the date of the meeting or any adjournment, postponement
or rescheduling thereof, and such update shall be delivered in writing to the secretary at the principal executive offices of the Corporation
no later than two (2) business days after the occurrence of any material change to the information previously disclosed pursuant
to Article I, Section 2(a)(2)(C).
(4) Notwithstanding
anything in the second sentence of Article I, Section 2(a)(2) of this Bylaw to the contrary, in the event that the number
of directors to be elected to the Board of Directors is increased and there is no public announcement naming all of the nominees for
director or specifying the size of the increased Board of Directors made by the Corporation at least ten (10) days before the last
day a stockholder may deliver a notice of nomination in accordance with the second sentence of Article I, Section 2(a)(2),
a stockholder’s notice required by this Bylaw shall also be considered timely, but only with respect to nominees for any new positions
created by such increase, if it shall be received by the Secretary of the Corporation not later than 5:00 p.m. Eastern time on the
tenth (10th) day following the day on which such public announcement is first made by the Corporation.
(b) General.
(1) Only
such persons who are nominated in accordance with the provisions of this Bylaw shall be eligible for election and to serve as directors,
and only such business shall be conducted at an Annual Meeting as shall have been brought before the meeting in accordance with the provisions
of this Bylaw or in accordance with Rule 14a-8 under the Exchange Act. The Board of Directors or a designated committee thereof
shall have the power to determine whether a nomination or any business proposed to be brought before the meeting was made in accordance
with the provisions of this Bylaw. If neither the Board of Directors nor such designated committee makes a determination as to whether
any stockholder proposal or nomination was made in accordance with the provisions of this Bylaw, the chair of the meeting (as defined
in Section 9 of this Article I) shall have the power and duty to determine whether the stockholder proposal or nomination was
made in accordance with the provisions of this Bylaw. If the Board of Directors or a designated committee thereof or the chair of the
meeting, as applicable, determines that any stockholder proposal or nomination was not made in accordance with the provisions of this
Bylaw, such proposal or nomination shall be disregarded and shall not be presented for action at the Annual Meeting.
(2) Except
as otherwise required by law, nothing in this Article I, Section 2 shall obligate the Corporation or the Board of Directors
to include in any proxy statement or other stockholder communication distributed on behalf of the Corporation or the Board of Directors
information with respect to any nominee for director or any other matter of business submitted by a stockholder.
(3) Notwithstanding
the foregoing provisions of this Article I, Section 2, if the nominating or proposing stockholder (or a qualified representative
of the stockholder) does not appear at the Annual Meeting to present a nomination or any business, such nomination or business shall
be disregarded, notwithstanding that proxies in respect of such vote may have been received by the Corporation. For purposes of this
Article I, Section 2, to be considered a qualified representative of the proposing stockholder, a person must be authorized
by a written instrument executed by such stockholder or an electronic transmission delivered by such stockholder to act for such stockholder
as proxy at the meeting of stockholders, and such person must produce such written instrument or electronic transmission, or a reliable
reproduction of the written instrument or electronic transmission, to the chair of the meeting at the meeting of stockholders.
(4) For
purposes of this Bylaw, “public announcement” shall mean disclosure in a press release reported by the Dow Jones News Service,
Associated Press or comparable national news service or in a document publicly filed by the Corporation with the Securities and Exchange
Commission pursuant to Section 13, 14 or 15(d) of the Exchange Act.
(5) Notwithstanding
the foregoing provisions of this Bylaw, a stockholder shall also comply with all applicable requirements of the Exchange Act and the
rules and regulations thereunder, including, but not limited to, Rule 14a-19 of the Exchange Act, with respect to the matters
set forth in this Bylaw. If a stockholder fails to comply with any applicable requirements of the Exchange Act, including, but not limited
to, Rule 14a-19 promulgated thereunder, such stockholder’s proposed nomination or proposed business shall be deemed to have
not been made in compliance with this Bylaw and shall be disregarded.
(6) Further
notwithstanding the foregoing provisions of this Bylaw, unless otherwise required by law, (i) no Proposing Person shall solicit
proxies in support of director nominees other than the Corporation’s nominees unless such Proposing Person has complied with Rule 14a-19
promulgated under the Exchange Act in connection with the solicitation of such proxies, including the provision to the Corporation of
notices required thereunder with timely notice and (ii) if any Proposing Person (A) provides notice pursuant to Rule 14a-19(b) promulgated
under the Exchange Act, (B) subsequently fails to comply with the requirements of Rule 14a-19(a)(2) or Rule 14a-19(a)(3) promulgated
under the Exchange Act, including the provision to the Corporation of notices required thereunder with timely notice and (C) no
other Proposing Person has provided notice pursuant to, and in compliance with, Rule 14a-19 under the Exchange Act that it intends
to solicit proxies in support of the election of such proposed nominee in accordance with Rule 14a-19(b) under the Exchange
Act, then such proposed nominee shall be disqualified from nomination, the Corporation shall disregard the nomination of such proposed
nominee and no vote on the election of such proposed nominee shall occur. Upon request by the Corporation, if any Proposing Person provides
notice pursuant to Rule 14a-19(b) promulgated under the Exchange Act, such Proposing Person shall deliver to the Corporation,
no later than five (5) business days prior to the applicable meeting date, reasonable evidence that it has met the requirements
of Rule 14a-19(a)(3) promulgated under the Exchange Act.
(7) The
number of nominees a stockholder may nominate for election at the Annual Meeting (or in the case of a stockholder giving the notice on
behalf of a beneficial owner, the number of nominees a stockholder may nominate for election at the Annual Meeting on behalf of such
beneficial owner) shall not exceed the number of directors to be elected at such Annual Meeting. A stockholder may not designate any
substitute nominees unless the stockholder provides timely notice of such substitute nominee(s) in accordance with these By-laws
(and such notice contains all of the information, representations, questionnaires and certifications with respect to such substitute
nominee(s) that are required by the By-laws with respect to nominees for director).
Section 3. Special
Meetings. Except as otherwise required by statute and subject to the rights, if any, of the holders of any series of Preferred Stock,
special meetings of the stockholders of the Corporation may be called only by or at the direction of the Board of Directors. The Board
of Directors may postpone or reschedule any previously scheduled special meeting of stockholders. Only those matters set forth in the
notice of the special meeting may be considered or acted upon at a special meeting of stockholders of the Corporation. Nominations of
persons for election to the Board of Directors and stockholder proposals of other business shall not be brought before a special meeting
of stockholders to be considered by the stockholders unless such special meeting is held in lieu of an annual meeting of stockholders
in accordance with Article I, Section 1 of these Bylaws, in which case such special meeting in lieu thereof shall be deemed
an Annual Meeting for purposes of these Bylaws and the provisions of Article I, Section 2 of these Bylaws shall govern such
special meeting.
Section 4. Notice
of Meetings; Adjournments.
(a) A
notice of each Annual Meeting stating the hour, date and place, if any, of such Annual Meeting, the means of remote communication, if
any, by which stockholders and proxyholders may be deemed to be present in person and vote at such meeting, and the record date for determining
the stockholders entitled to vote at the meeting, if such date is different from the record date for determining stockholders entitled
to notice of the meeting, shall be given not less than ten (10) days nor more than sixty (60) days before the Annual Meeting, to
each stockholder entitled to vote thereat by delivering such notice to such stockholder or by mailing it, postage prepaid, addressed
to such stockholder at the address of such stockholder as it appears on the Corporation’s stock transfer books. Without limiting
the manner by which notice may otherwise be given to stockholders, any notice to stockholders may be given by electronic transmission
in the manner provided in Section 232 of the DGCL.
(b) Notice
of all special meetings of stockholders shall be given in the same manner as provided for Annual Meetings, except that the notice of
all special meetings shall also state the purpose or purposes for which the meeting has been called.
(c) Notice
of an Annual Meeting or special meeting of stockholders need not be given to a stockholder if a waiver of notice is executed, or waiver
of notice by electronic transmission is provided, before or after such meeting by such stockholder or if such stockholder attends such
meeting, unless such attendance is for the express purpose of objecting at the beginning of the meeting to the transaction of any business
because the meeting was not lawfully called or convened.
(d) The
Board of Directors may postpone and reschedule or cancel any previously scheduled Annual Meeting or special meeting of stockholders and
any record date with respect thereto, regardless of whether any notice or public disclosure with respect to any such meeting has been
sent or made pursuant to Section 2 of this Article I or otherwise. In no event shall the public announcement of an adjournment,
postponement or rescheduling of any previously scheduled meeting of stockholders commence a new time period for the giving of a stockholder’s
notice under this Article I.
(e) When
any meeting is convened, the chair of the meeting or the stockholders present or represented by proxy at such meeting may adjourn the
meeting from time to time for any reason, regardless of whether a quorum is present, to reconvene at any other time and at any place
at which a meeting of stockholders may be held under these Bylaws. When any Annual Meeting or special meeting of stockholders is adjourned
to another hour, date or place (including an adjournment taken to address a technical failure to convene or continue a meeting using
remote communication), notice need not be given of the adjourned meeting if the time, place, if any, thereof and the means of remote
communications, if any, by which stockholders and proxy holders may be deemed to be present in person and vote at such adjourned meeting
are (i) announced at the meeting at which the adjournment is taken, (ii) displayed, during the time scheduled for the meeting,
on the same electronic network used to enable stockholders and proxy holders to participate in the meeting by means of remote communication
or (iii) set forth in the notice of meeting given in accordance with this Section 4; provided, however, that if the adjournment
is for more than thirty (30) days from the meeting date, or if after the adjournment a new record date is fixed for the adjourned meeting,
notice of the adjourned meeting and the means of remote communications, if any, by which stockholders and proxyholders may be deemed
to be present in person and vote at such adjourned meeting shall be given to each stockholder of record entitled to vote thereat and
each stockholder who, by law or under the Certificate of Incorporation of the Corporation (as the same may hereafter be amended and/or
restated, the “Certificate”) or these Bylaws, is entitled to such notice.
Section 5. Quorum.
Except as otherwise provided by law, the certificate of incorporation or these Bylaws, at each meeting of stockholders, the presence
in person or by remote communication, if applicable, or represented by proxy, of the holders of a majority in voting power of the outstanding
shares of stock entitled to vote at the meeting shall be necessary and sufficient to constitute a quorum. If less than a quorum is present
at a meeting, the chair of the meeting or the holders of voting stock, by the affirmative vote of a majority of the voting power present
in person or by proxy and entitled to vote thereon, may adjourn the meeting from time to time, and the meeting may be held as adjourned
without further notice, except as otherwise provided in Section 4 of this Article I. At such adjourned meeting at which a quorum
is present, any business may be transacted which might have been transacted at the meeting as originally noticed. The stockholders present
at a duly constituted meeting may continue to transact business until adjournment, notwithstanding the withdrawal of enough stockholders
to leave less than a quorum.
Section 6. Voting
and Proxies.
(a) The
stockholders entitled to vote at any meeting of stockholders shall be determined in accordance with the provisions of Article IV,
Section 4 of these Bylaws, subject to Section 217 (relating to voting rights of fiduciaries, pledgors and joint owners of stock)
and Section 218 (relating to voting trusts and other voting agreements) of the DGCL. Stockholders shall have one vote for each share
of stock entitled to vote owned by them of record according to the stock ledger of the Corporation as of the record date, unless otherwise
provided by law or by the Certificate. Stockholders may vote either (i) in person, (ii) by written proxy or (iii) by a
transmission permitted by Section 212(c) of the DGCL. Any copy, facsimile telecommunication or other reliable reproduction
of the writing or transmission permitted by Section 212(c) of the DGCL may be substituted for or used in lieu of the original
writing or transmission for any and all purposes for which the original writing or transmission could be used, provided that such copy,
facsimile telecommunication or other reproduction shall be a complete reproduction of the entire original writing or transmission. Proxies
shall be filed in accordance with the procedures established for the meeting of stockholders. A proxy with respect to stock held in the
name of two or more persons shall be valid if executed by or on behalf of any one of them unless at or prior to the exercise of the proxy
the Corporation receives a specific written notice to the contrary from any one of them. In the event the Corporation receives proxies
for disqualified or withdrawn nominees for the Board of Directors, such votes for such disqualified or withdrawn nominees in the proxies
will be treated as abstentions.
(b) Any
stockholder directly or indirectly soliciting proxies from other stockholders must use a proxy card color other than white, which shall
be reserved for the exclusive use by the Board of Directors.
Section 7. Action
at Meeting. When a quorum is present at any meeting of stockholders, any matter before any such meeting (other than an election of
a director or directors) shall be decided by a majority of the votes properly cast for and against such matter, except where a larger
vote is required by law, by the Certificate or by these Bylaws. Any election of directors by stockholders shall be determined by a plurality
of the votes properly cast on the election of directors.
Section 8. Stockholder
Lists. The Corporation shall prepare, no later than the tenth (10th) day before each Annual Meeting or special meeting
of stockholders, a complete list of the stockholders entitled to vote at the meeting, arranged in alphabetical order, and showing the
address of each stockholder and the number of shares registered in the name of each stockholder; provided, however, that if the record
date for determining the stockholders entitled to vote is less than ten (10) days before the meeting date, the list shall reflect
the stockholders entitled to vote as of the tenth (10th) day before the meeting date. Such list shall be open to the examination of any
stockholder for any purpose germane to the meeting for a period of ten (10) days ending on the day before the meeting date in the
manner provided by law.
Section 9. Conduct
of Meeting. The Board of Directors may adopt by resolution such rules, regulations and procedures for the conduct of any meeting
of stockholders as it shall deem appropriate. Except to the extent inconsistent with rules, regulations and procedures adopted by the
Board of Directors, the chair of the meeting shall have the right to prescribe such rules, regulations and procedures and to do all such
acts, as, in the judgment of such chair, are necessary, appropriate or convenient for the proper conduct of the meeting. Such rules,
regulations or procedures, whether adopted by the Board of Directors or the chair of the meeting, may include, without limitation, the
following: (a) the establishment of an agenda for the meeting; (b) rules and procedures for maintaining order at the meeting
and the safety of those present at the meeting; (c) limitations on attendance at or participation in the meeting to stockholders
of record of the Corporation, their duly authorized and constituted proxies, or such other persons as the chair of the meeting shall
determine; (d) restrictions on entry to the meeting after the time fixed for the commencement thereof; (e) the determination
of the circumstances in which any person may make a statement or ask questions and limitations on the time allotted to questions or comments;
(f) the determination of when the polls shall open and close for any given matter to be voted on at the meeting; (g) the exclusion
or removal of any stockholders or any other individual who refuses to comply with meeting rules, regulations, or procedures; (h) restrictions
on the use of audio and video recording devices, cell phones and other electronic devices; (i) rules, regulations and procedures
for compliance with any federal, state or local laws or regulations (including those concerning safety, health or security); (j) procedures
(if any) requiring attendees to provide the Corporation advance notice of their intent to attend the meeting; and (k) rules, regulations
or procedures regarding the participation by means of remote communication of stockholders and proxy holders not physically present at
a meeting, whether such meeting is to be held at a designated place or solely by means of remote communication. The chair of the meeting
shall be: (i) such person as the Board of Directors shall have designated to preside over all meetings of the stockholders; (ii) if
the Board of Directors has not so designated such a chair of the meeting or if the chair of the meeting is unable to so preside or is
absent, then the Chairperson of the Board, if one is elected; (iii) if the Board of Directors has not so designated a chair of the
meeting and there is no Chairperson of the Board, or if the chair of the meeting or the Chairperson of the Board is unable to so preside
or is absent, then the Chief Executive Officer, if one is elected; or (iv) in the absence or inability to serve of any of the aforementioned
persons, the President of the Corporation. Unless and to the extent determined by the Board of Directors or the chair of the meeting,
the chair of the meeting shall not be obligated to adopt or follow any technical, formal or parliamentary rules or principles of
procedure. In the absence of the Secretary of the Corporation, the secretary of the meeting shall be such person as the chair of the
meeting appoints.
Section 10. Inspectors
of Elections. The Corporation shall, in advance of any meeting of stockholders, appoint one or three inspectors to act at the meeting
and make a written report thereof. The Corporation may designate one or more persons as alternate inspectors to replace any inspector
who fails to act. If no inspector or alternate is able to act at a meeting of stockholders, the chair of the meeting officer shall appoint
one or more inspectors to act at the meeting. Any inspector may, but need not, be an officer, employee or agent of the Corporation. Each
inspector, before entering upon the discharge of his or her duties, shall take and sign an oath faithfully to execute the duties of inspector
with strict impartiality and according to the best of his or her ability. The inspectors shall perform such duties as are required by
the DGCL, including the counting of all votes and ballots. The inspectors may appoint or retain other persons or entities to assist the
inspectors in the performance of the duties of the inspectors. The chair of the meeting may review all determinations made by the inspectors,
and in so doing the chair of the meeting shall be entitled to exercise his or her sole judgment and discretion and he or she shall not
be bound by any determinations made by the inspectors. All determinations by the inspectors and, if applicable, the chair of the meeting,
shall be subject to further review by any court of competent jurisdiction.
Article II
Directors
Section 1. Powers.
The business and affairs of the Corporation shall be managed by or under the direction of the Board of Directors, except as otherwise
provided by the Certificate or required by law.
Section 2. Number
and Terms. The number of directors of the Corporation shall be fixed solely and exclusively by resolution duly adopted from time
to time by the Board of Directors, provided the Board of Directors shall consist of at least one (1) member. The directors shall
hold office in the manner provided in the Certificate.
Section 3. Qualification.
No director need be a stockholder of the Corporation.
Section 4. Vacancies.
Vacancies in the Board of Directors shall be filled in the manner provided in the Certificate.
Section 5. Removal.
Directors may be removed from office only in the manner provided in the Certificate or by applicable law.
Section 6. Resignation.
A director may resign at any time by electronic transmission or by giving written notice to the Chairperson of the Board, if one is elected,
the President or the Secretary. A resignation shall be effective upon receipt, unless the resignation otherwise provides.
Section 7. Regular
Meetings. Regular meetings of the Board of Directors may be held at such hour, date and place (if any) as the Board of Directors
may from time to time determine and publicize by means of reasonable notice given to any director who is not present when such determination
is made.
Section 8. Special
Meetings. Special meetings of the Board of Directors may be called, orally or in writing, by or at the request of a majority of the
directors, the Chairperson of the Board, if one is elected, or the President. The person calling any such special meeting of the Board
of Directors may fix the hour, date and place (if any) thereof. Notice thereof shall be given to each director as provided in Section 9
of this Article II.
Section 9. Notice
of Meetings. Notice of the hour, date and place (if any) of all special meetings of the Board of Directors shall be given to each
director by the Secretary or an Assistant Secretary, or in case of the death, absence, incapacity or refusal of such persons, by the
Chairperson of the Board, if one is elected, the President or such other officer designated by the Chairperson of the Board, if one is
elected, or any one of the directors calling the meeting. Notice of any special meeting of the Board of Directors shall be given to each
director in person, by telephone, or by facsimile, electronic mail or other form of electronic communication, sent to his or her business
or home address, at least twenty-four (24) hours in advance of the meeting, or by written notice mailed to his or her business or home
address, at least forty-eight (48) hours in advance of the meeting provided, however, that if the person or persons calling the meeting
determine that it is otherwise necessary or advisable to hold the meeting sooner, then such person or persons may prescribe a shorter
time period for notice to be given personally or by telephone, facsimile, electronic mail or other similar means of communication. Such
notice shall be deemed to be delivered when hand-delivered to such address; read to such director by telephone; deposited in the mail
so addressed, with postage thereon prepaid, if mailed; or dispatched or transmitted if sent by facsimile transmission or by electronic
mail or other form of electronic communication. A written waiver of notice signed or electronically transmitted before or after a meeting
by a director and filed with the records of the meeting shall be deemed to be equivalent to notice of the meeting. The attendance of
a director at a meeting shall constitute a waiver of notice of such meeting, except where a director attends a meeting for the express
purpose of objecting at the beginning of the meeting to the transaction of any business because such meeting is not lawfully called or
convened. Except as otherwise required by law, by the Certificate or by these Bylaws, neither the business to be transacted at, nor the
purpose of, any meeting of the Board of Directors need be specified in the notice or waiver of notice of such meeting.
Section 10. Quorum.
At any meeting of the Board of Directors, a majority of the total number of directors shall constitute a quorum for the transaction of
business, but if less than a quorum is present at a meeting, a majority of the directors present may adjourn the meeting from time to
time, and the meeting may be held as adjourned without further notice. Any business that might have been transacted at the meeting as
originally noticed may be transacted at such adjourned meeting at which a quorum is present. For purposes of this Article II, the
total number of directors includes any unfilled vacancies on the Board of Directors.
Section 11. Action
at Meeting. At any meeting of the Board of Directors at which a quorum is present, the affirmative vote of a majority of the directors
present shall constitute action by the Board of Directors, unless otherwise required by law, by the Certificate or by these Bylaws.
Section 12. Action
by Consent. Any action required or permitted to be taken at any meeting of the Board of Directors may be taken without a meeting
if all members of the Board of Directors consent thereto in writing or by electronic transmission. After such action is taken, the writing
or writings or electronic transmission or transmissions shall be filed with the records of the meetings of the Board of Directors. Such
filing shall be in paper form if the minutes are maintained in paper form and shall be in electronic form if the minutes are maintained
in electronic form. Such consent shall be treated as a resolution of the Board of Directors for all purposes.
Section 13. Manner
of Participation. Directors may participate in meetings of the Board of Directors by means of video conference, conference telephone
or other communications equipment by means of which all directors participating in the meeting can hear each other, and participation
in a meeting in accordance herewith shall constitute presence in person at such meeting for purposes of these Bylaws.
Section 14. Presiding
Director. The Board of Directors shall designate a representative to preside over all meetings of the Board of Directors, provided
that if the Board of Directors does not so designate such a presiding director or such designated presiding director is unable to so
preside or is absent, then the Chairperson of the Board, if one is elected, shall preside over all meetings of the Board of Directors.
If both the designated presiding director, if one is so designated, and the Chairperson of the Board, if one is elected, are unable to
preside or are absent, the Board of Directors shall designate an alternate representative to preside over a meeting of the Board of Directors.
Section 15. Committees.
The Board of Directors may designate one or more committees, including, without limitation, a Culture and Compensation Committee, a Nominating &
Corporate Governance Committee and an Audit Committee, and may delegate thereto some or all of its powers to such committee(s) except
those which by law, by the Certificate or by these Bylaws may not be delegated. Except as the Board of Directors may otherwise determine,
any such committee may make rules for the conduct of its business, but unless otherwise provided by the Board of Directors or in
such rules, its business shall be conducted so far as possible in the same manner as is provided by these Bylaws for the Board of Directors.
All members of such committees shall hold such offices at the pleasure of the Board of Directors. The Board of Directors may abolish
any such committee at any time. Any committee to which the Board of Directors delegates any of its powers or duties shall keep records
of its meetings.
Section 16. Compensation
of Directors. Directors shall receive such compensation for their services as shall be determined by the Board of Directors, or a
designated committee thereof, provided that directors who are serving the Corporation as employees shall not receive any salary or other
compensation for their services as directors of the Corporation.
Section 17. Emergency
By-laws. In the event of any emergency, disaster, catastrophe or other similar emergency condition of a type described in Section 110(a) of
the DGCL (an “Emergency”), notwithstanding any different or conflicting provisions in the DGCL, the Certificate or
these By-laws, during such Emergency:
(a) A
meeting of the Board of Directors or a committee thereof may be called by any director, the Chairperson of the Board, the Chief Executive
Officer, the President or the Secretary by such means as, in the judgment of the person calling the meeting, may be feasible at the time,
and notice of any such meeting of the Board of Directors or any committee may be given, in the judgment of the person calling the meeting,
only to such directors as it may be feasible to reach at the time and by such means as may be feasible at the time. Such notice shall
be given at such time in advance of the meeting as, in the judgment of the person calling the meeting, circumstances permit.
(b) The
director or directors in attendance at a meeting called in accordance with Section 17(a) of this Article II shall constitute
a quorum.
(c) No
officer, director or employee acting in accordance with this Section 17 shall be liable except for willful misconduct. No amendment,
repeal or change to this Section 17 shall modify the prior sentence with regard to actions taken prior to the time of such amendment,
repeal or change.
Article III
Officers
Section 1. Enumeration.
The officers of the Corporation shall consist of a President, a Treasurer, a Secretary and such other officers, including, without limitation,
a Chairperson of the Board, a Chief Executive Officer and one or more Vice Presidents (including Executive Vice Presidents or Senior
Vice Presidents), Assistant Vice Presidents, Assistant Treasurers and Assistant Secretaries, as the Board of Directors may determine.
Any number of offices may be held by the same person. The salaries and other compensation of the officers of the Corporation will be
fixed by or in the manner designated by the Board of Directors or a committee thereof to which the Board of Directors has delegated such
responsibility.
Section 2. Election.
The Board of Directors shall elect the President, the Treasurer and the Secretary. Other officers may be elected by the Board of Directors
or by such officers delegated such authority by the Board of Directors.
Section 3. Qualification.
No officer need be a stockholder or a director.
Section 4. Tenure.
Except as otherwise provided by the Certificate or by these Bylaws, each of the officers of the Corporation shall hold office until his
or her successor is elected and qualified or until his or her earlier death, resignation or removal.
Section 5. Resignation
and Removal. Any officer may resign by delivering his or her written or electronically transmitted resignation to the Corporation
addressed to the President or the Secretary, and such resignation shall be effective upon receipt, unless the resignation otherwise provides.
Any resignation is without prejudice to the rights, if any, of the Corporation under any contract to which the officer is a party. Except
as otherwise provided by law or by resolution of the Board of Directors, the Board of Directors may remove any officer. Except as the
Board of Directors may otherwise determine, no officer who resigns or is removed shall have any right to any compensation as an officer
for any period following his or her resignation or removal, or any right to damages on account of such removal, whether his or her compensation
be by the month or by the year or otherwise, unless such compensation is expressly provided in a duly authorized written agreement with
the Corporation.
Section 6. Absence
or Disability. In the event of the absence or disability of any officer, the Board of Directors may designate another officer to
act temporarily in place of such absent or disabled officer.
Section 7. Vacancies.
Any vacancy in any office may be filled for the unexpired portion of the term by the Board of Directors.
Section 8. President.
The President shall, subject to the direction of the Board of Directors, have such powers and shall perform such duties as the Board
of Directors may from time to time designate.
Section 9. Chairperson
of the Board. The Chairperson of the Board, if one is elected, shall have such powers and shall perform such duties as the Board
of Directors may from time to time designate.
Section 10. Chief
Executive Officer. The Chief Executive Officer, if one is elected, shall have such powers and shall perform such duties as the Board
of Directors may from time to time designate.
Section 11. Vice
Presidents and Assistant Vice Presidents. Any Vice President (including any Executive Vice President or Senior Vice President) and
any Assistant Vice President shall have such powers and shall perform such duties as the Board of Directors or the Chief Executive Officer
may from time to time designate.
Section 12. Treasurer
and Assistant Treasurers. The Treasurer shall, subject to the direction of the Board of Directors and except as the Board of Directors
or the Chief Executive Officer may otherwise provide, have general charge of the financial affairs of the Corporation and shall cause
to be kept accurate books of account. The Treasurer shall have custody of all funds, securities and valuable documents of the Corporation.
He or she shall have such other duties and powers as may be designated from time to time by the Board of Directors or the Chief Executive
Officer. Any Assistant Treasurer shall have such powers and perform such duties as the Board of Directors or the Chief Executive Officer
may from time to time designate.
Section 13. Secretary
and Assistant Secretaries. The Secretary shall record all the proceedings of the meetings of the stockholders and the Board of Directors
(including committees of the Board of Directors) in books kept for that purpose. In his or her absence from any such meeting, a temporary
secretary chosen at the meeting shall record the proceedings thereof. The Secretary shall have charge of the stock ledger (which may,
however, be kept by any transfer or other agent of the Corporation). The Secretary shall have custody of the seal of the Corporation,
and the Secretary or an Assistant Secretary shall have authority to affix it to any instrument requiring it, and, when so affixed, the
seal may be attested by his or her signature or that of an Assistant Secretary. The Secretary shall have such other duties and powers
as may be designated from time to time by the Board of Directors or the Chief Executive Officer. In the absence of the Secretary, any
Assistant Secretary may perform his or her duties and responsibilities. Any Assistant Secretary shall have such powers and perform such
duties as the Board of Directors or the Chief Executive Officer may from time to time designate.
Section 14. Other
Powers and Duties. Subject to these Bylaws and to such limitations as the Board of Directors may from time to time prescribe, the
officers of the Corporation shall each have such powers and duties as generally pertain to their respective offices, as well as such
powers and duties as from time to time may be conferred by the Board of Directors or the Chief Executive Officer.
Section 15. Representation
of Shares of Other Corporations. The Chairperson of the Board, the President, any Vice President, the Treasurer, the Secretary or
Assistant Secretary of this Corporation, or any other person authorized by the Board of Directors or the President or a Vice President,
is authorized to vote, represent and exercise on behalf of this Corporation all rights incident to any and all securities of any other
entity or entities standing in the name of this Corporation. The authority granted herein may be exercised either by such person directly
or by any other person authorized to do so by proxy or power of attorney duly executed by such person having the authority.
Section 16. Bonded
Officers. The Board of Directors may require any officer to give the Corporation a bond in such sum and with such surety or sureties
as shall be satisfactory to the Board of Directors upon such terms and conditions as the Board of Directors may specify, including, without
limitation, a bond for the faithful performance of his or her duties and for the restoration to the Corporation of all property in his
or her possession or under his or her control belonging to the Corporation.
Article IV
Capital Stock
Section 1. Certificates
of Stock. Each stockholder shall be entitled to a certificate of the capital stock of the Corporation in such form as may from time
to time be prescribed by the Board of Directors. Such certificate shall be signed by any two authorized officers of the Corporation.
The Corporation seal and the signatures by the Corporation’s officers, the transfer agent or the registrar may be facsimiles. In
case any officer, transfer agent or registrar who has signed or whose facsimile signature has been placed on such certificate shall have
ceased to be such officer, transfer agent or registrar before such certificate is issued, it may be issued by the Corporation with the
same effect as if he or she were such officer, transfer agent or registrar at the time of its issue. Every certificate for shares of
stock which are subject to any restriction on transfer and every certificate issued when the Corporation is authorized to issue more
than one class or series of stock shall contain such legend with respect thereto as is required by law. Notwithstanding anything to the
contrary provided in these Bylaws, the Board of Directors may provide by resolution or resolutions that some or all of any or all classes
or series of its stock shall be uncertificated shares (except that the foregoing shall not apply to shares represented by a certificate
until such certificate is surrendered to the Corporation), and by the approval and adoption of these Bylaws, the Board of Directors has
determined that all classes or series of the Corporation’s stock may be uncertificated, whether upon original issuance, re-issuance
or subsequent transfer.
Section 2. Transfers.
Subject to any restrictions on transfer and unless otherwise provided by the Board of Directors, shares of stock that are represented
by a certificate may be transferred on the books of the Corporation by the surrender to the Corporation or its transfer agent of the
certificate therefor properly endorsed or accompanied by a written assignment or power of attorney properly executed, with transfer stamps
(if necessary) affixed, and with such proof of the authenticity of signature as the Corporation or its transfer agent may reasonably
require. Shares of stock that are not represented by a certificate may be transferred on the books of the Corporation by submitting to
the Corporation or its transfer agent such evidence of transfer and following such other procedures as the Corporation or its transfer
agent may require.
Section 3. Stock
Transfer Agreements. The Corporation shall have power to enter into and perform any agreement with any number of stockholders of
any one or more classes of stock of the Corporation to restrict the transfer of shares of stock of the Corporation of any one or more
classes owned by such stockholders in any manner not prohibited by the DGCL.
Section 4. Record
Holders. Except as may otherwise be required by law, by the Certificate or by these Bylaws, the Corporation shall be entitled to
treat the record holder of stock as shown on its books as the owner of such stock for all purposes, including the payment of dividends
and the right to vote with respect thereto, regardless of any transfer, pledge or other disposition of such stock, until the shares have
been transferred on the books of the Corporation in accordance with the requirements of these Bylaws.
Section 5. Record
Date. In order that the Corporation may determine the stockholders entitled to notice of or to vote at any meeting of stockholders
or any adjournment thereof or entitled to receive payment of any dividend or other distribution or allotment of any rights, or entitled
to exercise any rights in respect of any change, conversion or exchange of stock or for the purpose of any other lawful action, the Board
of Directors may fix a record date, which record date shall not precede the date upon which the resolution fixing the record date is
adopted by the Board of Directors, and which record date: (a) in the case of determination of stockholders entitled to vote at any
meeting of stockholders, shall, unless otherwise required by law, not be more than sixty (60) nor less than ten (10) days before
the date of such meeting and (b) in the case of any other action, shall not be more than sixty (60) days prior to such other action.
If no record date is fixed: (i) the record date for determining stockholders entitled to notice of or to vote at a meeting of stockholders
shall be at 5:00 p.m. Eastern time on the day next preceding the day on which notice is given, or, if notice is waived, at 5:00
p.m. Eastern time on the day next preceding the day on which the meeting is held; and (ii) the record date for determining
stockholders for any other purpose shall be at 5:00 p.m. Eastern time on the day on which the Board of Directors adopts the resolution
relating thereto.
Section 6. Replacement
of Certificates. In case of the alleged loss, destruction or mutilation of a certificate of stock of the Corporation, a duplicate
certificate may be issued in place thereof, upon such terms as the Board of Directors may prescribe.
Article V
Indemnification
Section 1. Definitions.
For purposes of this Article V:
(a) “Corporate
Status” describes the status of a person who is serving or has served (i) as a Director of the Corporation, (ii) as an
Officer of the Corporation, (iii) as a Non-Officer Employee of the Corporation or (iv) as a director, partner, trustee, officer,
employee or agent of any other corporation, partnership, limited liability company, joint venture, trust, employee benefit plan, foundation,
association, organization or other legal entity which such person is or was serving at the request of the Corporation. For purposes of
this Section 1(a), a Director, Officer or Non-Officer Employee of the Corporation who is serving or has served as a director, partner,
trustee, officer, employee or agent of a Subsidiary shall be deemed to be serving at the request of the Corporation. Notwithstanding
the foregoing, “Corporate Status” shall not include the status of a person who is serving or has served as a director, officer,
employee or agent of a constituent corporation absorbed in a merger or consolidation transaction with the Corporation with respect to
such person’s activities prior to said transaction, unless specifically authorized by the Board of Directors or the stockholders
of the Corporation;
(b) “Director”
means any person who serves or has served the Corporation as a director on the Board of Directors of the Corporation;
(c) “Disinterested
Director” means, with respect to each Proceeding in respect of which indemnification is sought hereunder, a Director of the Corporation
who is not and was not a party to such Proceeding;
(d) “Expenses”
means all attorneys’ fees, retainers, court costs, transcript costs, fees of expert witnesses, private investigators and professional
advisors (including, without limitation, accountants and investment bankers), travel expenses, duplicating costs, printing and binding
costs, costs of preparation of demonstrative evidence and other courtroom presentation aids and devices, costs incurred in connection
with document review, organization, imaging and computerization, telephone charges, postage, delivery service fees, and all other disbursements,
costs or expenses of the type customarily incurred in connection with prosecuting, defending, preparing to prosecute or defend, investigating,
being or preparing to be a witness in, settling or otherwise participating in, a Proceeding;
(e) “Liabilities”
means judgments, damages, liabilities, losses, penalties, excise taxes, fines and amounts paid in settlement;
(f) “Non-Officer
Employee” means any person who serves or has served as an employee or agent of the Corporation, but who is not or was not a Director
or Officer;
(g) “Officer”
means any person who serves or has served the Corporation as an officer of the Corporation appointed by the Board of Directors of the
Corporation;
(h) “Proceeding”
means any threatened, pending or completed action, suit, arbitration, alternate dispute resolution mechanism, inquiry, investigation,
administrative hearing or other proceeding, whether civil, criminal, administrative, arbitrative or investigative; and
(i) “Subsidiary”
means any corporation, partnership, limited liability company, joint venture, trust or other entity of which the Corporation owns (either
directly or through or together with another Subsidiary of the Corporation) either (i) a general partner, managing member or other
similar interest or (ii) (A) fifty percent (50%) or more of the voting power of the voting capital equity interests of such
corporation, partnership, limited liability company, joint venture or other entity, or (B) fifty percent (50%) or more of the outstanding
voting capital stock or other voting equity interests of such corporation, partnership, limited liability company, joint venture or other
entity.
Section 2. Indemnification
of Directors and Officers.
(a) Subject
to the operation of Section 4 of this Article V, each Director and Officer shall be indemnified and held harmless by the Corporation
to the fullest extent authorized by the DGCL, as the same exists or may hereafter be amended (but, in the case of any such amendment,
only to the extent that such amendment permits the Corporation to provide broader indemnification rights than such law permitted the
Corporation to provide prior to such amendment), and to the extent authorized in this Section 2.
(1) Actions,
Suits and Proceedings Other than By or In the Right of the Corporation. Each Director and Officer shall be indemnified and held harmless
by the Corporation against any and all Expenses and Liabilities that are incurred or paid by such Director or Officer or on such Director’s
or Officer’s behalf in connection with any Proceeding or any claim, issue or matter therein (other than an action by or in the
right of the Corporation), which such Director or Officer is, or is threatened to be made, a party to or participant in by reason of
such Director’s or Officer’s Corporate Status, if such Director or Officer acted in good faith and in a manner such Director
or Officer reasonably believed to be in or not opposed to the best interests of the Corporation and, with respect to any criminal proceeding,
had no reasonable cause to believe his or her conduct was unlawful.
(2) Actions,
Suits and Proceedings By or In the Right of the Corporation. Each Director and Officer shall be indemnified and held harmless by
the Corporation against any and all Expenses that are incurred by such Director or Officer or on such Director’s or Officer’s
behalf in connection with any Proceeding or any claim, issue or matter therein by or in the right of the Corporation, which such Director
or Officer is, or is threatened to be made, a party to or participant in by reason of such Director’s or Officer’s Corporate
Status, if such Director or Officer acted in good faith and in a manner such Director or Officer reasonably believed to be in or not
opposed to the best interests of the Corporation; provided, however, that no indemnification shall be made under this Section 2(a)(2) in
respect of any claim, issue or matter as to which such Director or Officer shall have been finally adjudged by a court of competent jurisdiction
to be liable to the Corporation, unless, and only to the extent that, the Court of Chancery of the State of Delaware or another court
in which such Proceeding was brought shall determine upon application that, despite adjudication of liability, but in view of all the
circumstances of the case, such Director or Officer is fairly and reasonably entitled to indemnification for such Expenses that such
court deems proper.
(3) Survival
of Rights. The rights of indemnification provided by this Section 2 shall continue as to a Director or Officer after he or she
has ceased to be a Director or Officer and shall inure to the benefit of his or her heirs, executors, administrators and personal representatives.
(4) Actions
by Directors or Officers. Notwithstanding the foregoing, the Corporation shall indemnify any Director or Officer seeking indemnification
in connection with a Proceeding initiated by such Director or Officer only if such Proceeding (including any parts of such Proceeding
not initiated by such Director or Officer) was authorized in advance by the Board of Directors, unless such Proceeding was brought to
enforce such Officer’s or Director’s rights to indemnification or, in the case of Directors, advancement of Expenses under
these Bylaws in accordance with the provisions set forth herein.
Section 3. Indemnification
of Non-Officer Employees. Subject to the operation of Section 4 of this Article V, each Non-Officer Employee may, in the
discretion of the Board of Directors, be indemnified by the Corporation to the fullest extent authorized by the DGCL, as the same exists
or may hereafter be amended, against any or all Expenses and Liabilities that are incurred by such Non-Officer Employee or on such Non-Officer
Employee’s behalf in connection with any threatened, pending or completed Proceeding, or any claim, issue or matter therein, which
such Non-Officer Employee is, or is threatened to be made, a party to or participant in by reason of such Non-Officer Employee’s
Corporate Status, if such Non-Officer Employee acted in good faith and in a manner such Non-Officer Employee reasonably believed to be
in or not opposed to the best interests of the Corporation and, with respect to any criminal proceeding, had no reasonable cause to believe
his or her conduct was unlawful. The rights of indemnification provided by this Section 3 shall exist as to a Non-Officer Employee
after he or she has ceased to be a Non-Officer Employee and shall inure to the benefit of his or her heirs, personal representatives,
executors and administrators. Notwithstanding the foregoing, the Corporation may indemnify any Non-Officer Employee seeking indemnification
in connection with a Proceeding initiated by such Non-Officer Employee only if such Proceeding was authorized in advance by the Board
of Directors.
Section 4. Determination.
Unless ordered by a court, no indemnification shall be provided pursuant to this Article V to a Director, to an Officer or to a
Non-Officer Employee unless a determination shall have been made that such person acted in good faith and in a manner such person reasonably
believed to be in or not opposed to the best interests of the Corporation and, with respect to any criminal Proceeding, such person had
no reasonable cause to believe his or her conduct was unlawful. Such determination shall be made by (a) a majority vote of the Disinterested
Directors, even though less than a quorum of the Board of Directors, (b) a committee comprised of Disinterested Directors, such
committee having been designated by a majority vote of the Disinterested Directors (even though less than a quorum), (c) if there
are no such Disinterested Directors, or if a majority of Disinterested Directors so directs, by independent legal counsel in a written
opinion or (d) by the stockholders of the Corporation.
Section 5. Advancement
of Expenses to Directors Prior to Final Disposition.
(a) The
Corporation shall advance all Expenses incurred by or on behalf of any Director in connection with any Proceeding in which such Director
is involved by reason of such Director’s Corporate Status within thirty (30) days after the receipt by the Corporation of a written
statement from such Director requesting such advance or advances from time to time, whether prior to or after final disposition of such
Proceeding. Such statement or statements shall reasonably evidence the Expenses incurred by such Director and shall be preceded or accompanied
by an undertaking by or on behalf of such Director to repay any Expenses so advanced if it shall ultimately be determined that such Director
is not entitled to be indemnified against such Expenses. Notwithstanding the foregoing, the Corporation shall advance all Expenses incurred
by or on behalf of any Director seeking advancement of expenses hereunder in connection with a Proceeding initiated by such Director
only if such Proceeding (including any parts of such Proceeding not initiated by such Director) was (i) authorized by the Board
of Directors or (ii) brought to enforce such Director’s rights to indemnification or advancement of Expenses under these Bylaws.
(b) If
a claim for advancement of Expenses hereunder by a Director is not paid in full by the Corporation within thirty (30) days after receipt
by the Corporation of documentation of Expenses and the required undertaking, such Director may at any time thereafter bring suit against
the Corporation to recover the unpaid amount of the claim and, if successful in whole or in part, such Director shall also be entitled
to be paid the expenses of prosecuting such claim. The failure of the Corporation (including its Board of Directors or any committee
thereof, independent legal counsel or stockholders) to make a determination concerning the permissibility of such advancement of Expenses
under this Article V shall not be a defense to an action brought by a Director for recovery of the unpaid amount of an advancement
claim and shall not create a presumption that such advancement is not permissible. The burden of proving that a Director is not entitled
to an advancement of expenses shall be on the Corporation.
(c) In
any suit brought by the Corporation to recover an advancement of expenses pursuant to the terms of an undertaking, the Corporation shall
be entitled to recover such expenses upon a final adjudication that the Director has not met any applicable standard for indemnification
set forth in the DGCL.
Section 6. Advancement
of Expenses to Officers and Non-Officer Employees Prior to Final Disposition.
(a) The
Corporation may, at the discretion of the Board of Directors, advance any or all Expenses incurred by or on behalf of any Officer or
any Non-Officer Employee in connection with any Proceeding in which such person is involved by reason of his or her Corporate Status
as an Officer or Non-Officer Employee upon the receipt by the Corporation of a statement or statements from such Officer or Non-Officer
Employee requesting such advance or advances from time to time, whether prior to or after final disposition of such Proceeding. Such
statement or statements shall reasonably evidence the Expenses incurred by such Officer or Non-Officer Employee and shall be preceded
or accompanied by an undertaking by or on behalf of such person to repay any Expenses so advanced if it shall ultimately be determined
that such Officer or Non-Officer Employee is not entitled to be indemnified against such Expenses.
(b) In
any suit brought by the Corporation to recover an advancement of expenses pursuant to the terms of an undertaking, the Corporation shall
be entitled to recover such expenses upon a final adjudication that the Officer or Non-Officer Employee has not met any applicable standard
for indemnification set forth in the DGCL.
Section 7. Contractual
Nature of Rights.
(a) The
provisions of this Article V shall be deemed to be a contract between the Corporation and each Director and Officer entitled to
the benefits hereof at any time while this Article V is in effect, in consideration of such person’s past or current and any
future performance of services for the Corporation. Neither amendment, repeal or modification of any provision of this Article V
nor the adoption of any provision of the Certificate inconsistent with this Article V shall eliminate or reduce any right conferred
by this Article V in respect of any act or omission occurring, or any cause of action or claim that accrues or arises or any state
of facts existing, at the time of or before such amendment, repeal, modification or adoption of an inconsistent provision (even in the
case of a proceeding based on such a state of facts that is commenced after such time), and all rights to indemnification and advancement
of Expenses granted herein or arising out of any act or omission shall vest at the time of the act or omission in question, regardless
of when or if any proceeding with respect to such act or omission is commenced. The rights to indemnification and to advancement of expenses
provided by, or granted pursuant to, this Article V shall continue notwithstanding that the person has ceased to be a director or
officer of the Corporation and shall inure to the benefit of the estate, heirs, executors, administrators, legatees and distributees
of such person.
(b) If
a claim for indemnification hereunder by a Director or Officer is not paid in full by the Corporation within sixty (60) days after receipt
by the Corporation of a written claim for indemnification, such Director or Officer may at any time thereafter bring suit against the
Corporation to recover the unpaid amount of the claim, and if successful in whole or in part, such Director or Officer shall also be
entitled to be paid the expenses of prosecuting such claim. The failure of the Corporation (including its Board of Directors or any committee
thereof, independent legal counsel or stockholders) to make a determination concerning the permissibility of such indemnification under
this Article V shall not be a defense to an action brought by a Director or Officer for recovery of the unpaid amount of an indemnification
claim and shall not create a presumption that such indemnification is not permissible. The burden of proving that a Director or Officer
is not entitled to indemnification shall be on the Corporation.
(c) In
any suit brought by a Director or Officer to enforce a right to indemnification hereunder, it shall be a defense that such Director or
Officer has not met any applicable standard for indemnification set forth in the DGCL.
Section 8. Non-Exclusivity
of Rights. The rights to indemnification and to advancement of Expenses set forth in this Article V shall not be exclusive of
any other right that any Director, Officer or Non-Officer Employee may have or hereafter acquire under any statute, provision of the
Certificate or these Bylaws, agreement, vote of stockholders or Disinterested Directors or otherwise.
Section 9. Insurance.
The Corporation may maintain insurance, at its expense, to protect itself and any Director, Officer or Non-Officer Employee against any
liability of any character asserted against or incurred by the Corporation or any such Director, Officer or Non-Officer Employee, or
arising out of any such person’s Corporate Status, whether or not the Corporation would have the power to indemnify such person
against such liability under the DGCL or the provisions of this Article V.
Section 10. Other
Indemnification. The Corporation’s obligation, if any, to indemnify or provide advancement of Expenses to any person under
this Article V as a result of such person serving, at the request of the Corporation, as a director, partner, trustee, officer,
employee or agent of another corporation, partnership, joint venture, trust, employee benefit plan or other enterprise shall be reduced
by any amount such person may collect as indemnification or advancement of Expenses from such other corporation, partnership, joint venture,
trust, employee benefit plan or enterprise (the “Primary Indemnitor”). Any indemnification or advancement of Expenses
under this Article V owed by the Corporation as a result of a person serving, at the request of the Corporation, as a director,
partner, trustee, officer, employee or agent of another corporation, partnership, joint venture, trust, employee benefit plan or other
enterprise shall only be in excess of, and shall be secondary to, the indemnification or advancement of Expenses available from the applicable
Primary Indemnitor(s) and any applicable insurance policies.
Section 11. Savings
Clause. If this Article V or any portion hereof shall be invalidated on any ground by any court of competent jurisdiction, then
the Corporation shall nevertheless indemnify each indemnitee as to any expenses (including, without limitation, attorneys’ fees),
liabilities, losses, judgments, fines (including, without limitation, excise taxes and penalties arising under the Employee Retirement
Income Security Act of 1974, as amended) and amounts paid in settlement in connection with any action, suit, proceeding or investigation,
whether civil, criminal or administrative, including, without limitation, an action by or in the right of the Corporation, to the fullest
extent permitted by any applicable portion of this Article V that shall not have been invalidated and to the fullest extent permitted
by applicable law.
Article VI
Miscellaneous
Provisions
Section 1. Fiscal
Year. The fiscal year of the Corporation shall be determined by the Board of Directors.
Section 2. Seal.
The Board of Directors shall have power to adopt and alter the seal of the Corporation.
Section 3. Execution
of Instruments. All deeds, leases, transfers, contracts, bonds, notes and other obligations to be entered into by the Corporation
in the ordinary course of its business without director action may be executed on behalf of the Corporation by the Chairperson of the
Board, if one is elected, the President or the Treasurer or any other officer, employee or agent of the Corporation as the Board of Directors
or an executive committee of the Board of Directors may authorize or determine.
Section 4. Voting
of Securities. Unless the Board of Directors otherwise provides, the Chairperson of the Board, if one is elected, the President or
the Treasurer may waive notice of, and act on behalf of the Corporation, or appoint another person or persons to act as proxy or attorney
in fact for the Corporation with or without discretionary power and/or power of substitution, at any meeting of stockholders or stockholders
of any other corporation or organization, any of whose securities are held by the Corporation.
Section 5. Resident
Agent. The Board of Directors may appoint a resident agent upon whom legal process may be served in any action or proceeding against
the Corporation.
Section 6. Corporate
Records. The original or attested copies of the Certificate, Bylaws and records of all meetings of the incorporators, stockholders
and the Board of Directors and the stock transfer books, which shall contain the names of all stockholders, their record addresses and
the amount of stock held by each, may be kept outside the State of Delaware and shall be kept at the principal office of the Corporation,
at an office of its counsel, at an office of its transfer agent or in such manner as may be permitted by law.
Section 7. Certificate.
All references in these Bylaws to the Certificate shall be deemed to refer to the Certificate, as amended and/or restated and in effect
from time to time.
Section 8. Exclusive
Jurisdiction of Delaware Courts or the United States Federal District Courts. Unless the Corporation consents in writing to the selection
of an alternative forum, the Court of Chancery of the State of Delaware shall be the sole and exclusive forum for (i) any derivative
action or proceeding brought on behalf of the Corporation, (ii) any action asserting a claim of, or a claim based on, a breach of
a fiduciary duty owed by any current or former director, officer or other employee or stockholder of the Corporation to the Corporation
or the Corporation’s stockholders, (iii) any action asserting a claim arising pursuant to any provision of the DGCL or the
Certificate or these Bylaws (including the interpretation, validity or enforceability thereof) or as to which the DGCL confers jurisdiction
on the Court of Chancery of the State of Delaware or (iv) any action asserting a claim governed by the internal affairs doctrine;
provided, however, that this sentence will not apply to any causes of action arising under the Securities Act of 1933, as amended, or
the Exchange Act, or to any claim for which the federal courts have exclusive jurisdiction. Unless the Corporation consents in writing
to the selection of an alternative forum, the federal district courts of the United States of America shall be the sole and exclusive
forum for resolving any complaint asserting a cause of action arising under the Securities Act of 1933, as amended, the Exchange Act,
or the respective rules and regulations promulgated thereunder. To the fullest extent permitted by law, any person or entity purchasing
or otherwise acquiring any interest in shares of capital stock of the Corporation shall be deemed to have notice of and consented to
the provisions of this Section 8.
Section 9. Amendment
of Bylaws.
(a) Amendment
by Directors. Except as otherwise required by law, these Bylaws may be amended or repealed by the Board of Directors.
(b) Amendment
by Stockholders. Except as otherwise provided herein, the Bylaws of the Corporation may be amended or repealed at any annual meeting
of stockholders, or at any special meeting of stockholders called for such purpose, by the affirmative vote of the holders of not less
than two-thirds (2/3) of the voting power of the outstanding shares of capital stock entitled to vote on such amendment or repeal, voting
together as a single class; provided, however, that if the Board of Directors recommends that stockholders approve such amendment or
repeal at such meeting of stockholders, such amendment or repeal shall only require the affirmative vote of the majority of outstanding
shares of capital stock entitled to vote on such amendment or repeal, voting together as a single class.
Section 10. Notices.
If mailed, notice to stockholders shall be deemed given when deposited in the mail, postage prepaid, directed to the stockholder at such
stockholder’s address as it appears on the records of the Corporation. Without limiting the manner by which notice otherwise may
be given to stockholders, any notice to stockholders may be given by electronic transmission in the manner provided in Section 232
of the DGCL.
Section 11. Waivers.
A written waiver of any notice, signed by a stockholder or director, or waiver by electronic transmission by such person, whether given
before or after the time of the event for which notice is to be given, shall be deemed equivalent to the notice required to be given
to such person. Neither the business to be transacted at, nor the purpose of, any meeting need be specified in such a waiver.
Approved by the
Board of Directors on September 19, 2026, approved by the stockholders on October 8, 2026, and effective upon the effectiveness
of the Corporation’s Registration Statement on Form S-1 for its initial public offering.
Exhibit 5.1
October 9, 2026
City Therapeutics, Inc.
399 Binney Street
Cambridge, MA 02142
Re: Securities
Registered under Registration Statement on Form S-1
We have acted as counsel to you in connection with your filing of a
Registration Statement on Form S-1 (File No. 333-299117) (as amended or supplemented, the “Registration Statement”)
pursuant to the Securities Act of 1933, as amended (the “Securities Act”), relating to the registration of the offering by
City Therapeutics, Inc., a Delaware corporation (the “Company”), of up to 11,180,555 shares (the “Shares”)
of the Company’s common stock, $0.0001 par value per share, including Shares purchasable by the underwriters upon their exercise
of an over-allotment option granted to the underwriters by the Company. The Shares are being sold to the several underwriters named in,
and pursuant to, an underwriting agreement among the Company and such underwriters (the “Underwriting Agreement”).
We have reviewed such documents and made such examination of law as
we have deemed appropriate to give the opinions set forth below. We have relied, without independent verification, on certificates of
public officials and, as to matters of fact material to the opinions set forth below, on certificates of officers of the Company.
The opinion set forth below is limited to the Delaware General Corporation
Law.
Based on the foregoing, we are of the opinion that the Shares have
been duly authorized and, when delivered and paid for in accordance with the terms of the Underwriting Agreement, will be validly issued,
fully paid and non-assessable.
This opinion letter and the opinion it contains shall be interpreted
in accordance with the Core Opinion Principles as published in 74 Business Lawyer 815 (Summer 2019).
We hereby consent to the inclusion of this opinion as Exhibit 5.1
to the Registration Statement and to the references to our firm under the caption “Legal Matters” in the Registration Statement.
In giving our consent, we do not admit that we are in the category of persons whose consent is required under Section 7 of the Securities
Act or the rules and regulations thereunder.
| |
Very truly yours, |
| |
|
| |
/s/ Goodwin Procter LLP |
| |
|
| |
GOODWIN PROCTER LLP |
Exhibit 10.2
CITY THERAPEUTICS, INC.
2026 STOCK OPTION AND INCENTIVE PLAN
SECTION 1. GENERAL
PURPOSE OF THE PLAN; DEFINITIONS
The name of the plan is the
City Therapeutics, Inc. 2026 Stock Option and Incentive Plan (as amended from time to time, the “Plan”). The purpose
of the Plan is to encourage and enable the officers, employees, Non-Employee Directors and Consultants of City Therapeutics, Inc.
(the “Company”) and its Affiliates upon whose judgment, initiative and efforts the Company largely depends for the successful
conduct of its business to acquire a proprietary interest in the Company. It is anticipated that providing such persons with a direct
stake in the Company’s welfare will assure a closer identification of their interests with those of the Company and its stockholders,
thereby stimulating their efforts on the Company’s behalf and strengthening their desire to remain with the Company or one of its
Affiliates.
The following terms shall
be defined as set forth below:
“Act”
means the U.S. Securities Act of 1933, as amended, and the rules and regulations thereunder.
“Administrator”
means either the Board or the culture and compensation committee of the Board or a similar committee performing the functions of the culture
and compensation committee and which is comprised of not less than two Non-Employee Directors who are independent.
“Affiliate”
means, at the time of determination, any “parent” or “subsidiary” of the Company as such terms are defined in
Rule 405 of the Act. The Board will have the authority to determine the time or times at which “parent” or “subsidiary”
status is determined within the foregoing definition.
“Award”
or “Awards,” except where referring to a particular category of grant under the Plan, shall include Incentive Stock
Options, Non-Qualified Stock Options, Stock Appreciation Rights, Restricted Stock Units, Restricted Stock Awards, Unrestricted Stock Awards,
Cash-Based Awards and Dividend Equivalent Rights.
“Award
Agreement” means a written or electronic document setting forth the terms and provisions applicable to an Award granted
under the Plan. Each Award Agreement is subject to the terms and conditions of the Plan.
“Board”
means the Board of Directors of the Company.
“Cash-Based
Award” means an Award entitling the recipient to receive a cash-denominated payment.
“Code”
means the U.S. Internal Revenue Code of 1986, as amended, and any successor Code, and related rules, regulations and interpretations.
“Consultant”
means a consultant or adviser who provides bona fide services to the Company or an Affiliate as an independent contractor and who
qualifies as a consultant or advisor under Instruction A.1.(a)(1) of Form S-8 under the Act.
“Dividend
Equivalent Right” means an Award entitling the grantee to receive credits based on ordinary cash dividends that would
have been paid on the shares of Stock specified in the Dividend Equivalent Right (or other award to which it relates) if such shares had
been issued to and held by the grantee.
“Effective
Date” means the date on which the Plan becomes effective as set forth in Section 19.
“Exchange
Act” means the U.S. Securities Exchange Act of 1934, as amended, and the rules and regulations thereunder.
“Fair
Market Value” of the Stock on any given date means the fair market value of the Stock determined in good faith by the
Administrator; provided, however, that if the Stock is listed on the National Association of Securities Dealers Automated Quotation System
(“NASDAQ”), NASDAQ Global Market, The New York Stock Exchange or another national securities exchange or traded on any established
market, the determination shall be made by reference to the closing price. If there is no closing price for such date, the determination
shall be made by reference to the last date preceding such date for which there is a closing price; provided further, however, that if
the date for which Fair Market Value is determined is the Registration Date, the Fair Market Value shall be the “Price to the Public”
(or equivalent) set forth on the cover page for the final prospectus relating to the Company’s initial public offering.
“Incentive
Stock Option” means any Stock Option designated and qualified as an “incentive stock option” as defined in
Section 422 of the Code.
“Non-Employee
Director” means a member of the Board who is not also an employee of the Company or any Subsidiary.
“Non-Qualified
Stock Option” means any Stock Option that is not an Incentive Stock Option.
“Option”
or “Stock Option” means any option to purchase shares of Stock granted pursuant to Section 5.
“Outstanding
Shares” means, as of a specified date, the sum of (a) the number of shares of Stock issued and outstanding and (b) the
number of shares of Stock issuable pursuant to the exercise of any outstanding, pre-funded warrants to acquire Stock for a nominal exercise
price.
“Prior
Plan” means the Company’s 2023 Stock Plan, as amended.
“Registration
Date” means the date upon which the registration statement on Form S-1 that is filed by the Company with respect
to its initial public offering is declared effective by the U.S. Securities and Exchange Commission.
“Restricted
Shares” means the shares of Stock underlying a Restricted Stock Award that remain subject to a risk of forfeiture or
the Company’s right of repurchase.
“Restricted
Stock Award” means an Award of Restricted Shares subject to such restrictions and conditions as the Administrator may
determine at the time of grant.
“Restricted
Stock Units” means an Award of stock units subject to such restrictions and conditions as the Administrator may determine
at the time of grant.
“Sale
Event” shall mean (i) the sale of all or substantially all of the assets of the Company on a consolidated basis
to an unrelated person or entity, (ii) a merger, reorganization or consolidation pursuant to which the holders of the Company’s
outstanding voting power and outstanding stock immediately prior to such transaction do not own a majority of the outstanding voting power
and outstanding stock or other equity interests of the resulting or successor entity (or its ultimate parent, if applicable) immediately
upon completion of such transaction, (iii) the sale of all of the Stock of the Company to an unrelated person, entity or group thereof
acting in concert or (iv) any other transaction in which the owners of the Company’s outstanding voting power immediately prior
to such transaction do not own at least a majority of the outstanding voting power of the Company or any successor entity immediately
upon completion of the transaction other than as a result of the acquisition of securities directly from the Company. Notwithstanding
anything in the foregoing to the contrary, with respect to compensation (A) that is subject to Section 409A of the Code and
(B) for which a Sale Event would accelerate the timing of payment thereunder, the term “Sale Event” shall mean an event
that is both (I) a Sale Event (as defined above) and (II) a “change in control event” (within the meaning of Section 409A
of the Code).
“Sale Price”
means the value as determined by the Administrator of the consideration payable, or otherwise to be received by stockholders, per share
of Stock pursuant to a Sale Event.
“Section 409A”
means Section 409A of the Code and the regulations and other guidance promulgated thereunder.
“Service
Relationship” means any relationship as an employee, Non-Employee Director or Consultant of the Company or any Affiliate.
Unless as otherwise set forth in the Award Agreement, a Service Relationship shall be deemed to continue without interruption in the event
a grantee’s status changes from full-time employee to part-time employee or a grantee’s status changes from employee to Consultant
or Non-Employee Director or vice versa, provided that there is no interruption or other termination of Service Relationship in connection
with the grantee’s change in capacity.
“Stock”
means the Common Stock, par value $0.0001 per share, of the Company, subject to adjustments pursuant to Section 3.
“Stock
Appreciation Right” means an Award entitling the recipient to receive shares of Stock (or cash, to the extent explicitly
provided for in the applicable Award Agreement) having a value equal to the excess of the Fair Market Value of the Stock on the date of
exercise over the exercise price of the Stock Appreciation Right multiplied by the number of shares of Stock with respect to which the
Stock Appreciation Right shall have been exercised.
“Subsidiary”
means any corporation or other entity (other than the Company) in which the Company has at least a 50 percent interest, either directly
or indirectly.
“Substitute Awards”
means Awards granted or Stock issued by the Company in assumption of, or in substitution or exchange for, awards previously granted,
in each case by a company acquired by the Company or any Affiliate or with which the Company or any Affiliate combines.
“Ten
Percent Owner” means an employee who owns or is deemed to own (by reason of the attribution rules of Section 424(d) of
the Code) more than 10 percent of the combined voting power of all classes of stock of the Company or any parent or subsidiary corporation.
“Unrestricted
Stock Award” means an Award of shares of Stock free of any restrictions.
SECTION 2. ADMINISTRATION
OF PLAN; ADMINISTRATOR AUTHORITY TO SELECT GRANTEES AND DETERMINE AWARDS
(a) Administration
of Plan. The Plan shall be administered by the Administrator.
(b) Powers
of Administrator. The Administrator shall have the power and authority to grant Awards consistent with the terms of the Plan, including
the power and authority:
(i) to
select the individuals to whom Awards may from time to time be granted;
(ii) to
determine the time or times of grant, and the extent, if any, of Incentive Stock Options, Non-Qualified Stock Options, Stock Appreciation
Rights, Restricted Stock Awards, Restricted Stock Units, Unrestricted Stock Awards, Cash-Based Awards and Dividend Equivalent Rights,
or any combination of the foregoing, granted to any one or more grantees;
(iii) to
determine the number of shares of Stock to be covered by any Award;
(iv) to
determine and modify from time to time the terms and conditions, including restrictions, not inconsistent with the terms of the Plan,
of any Award, which terms and conditions may differ among individual Awards and grantees, and to approve the forms of Award Agreements;
(v) to
accelerate at any time the exercisability or vesting of all or any portion of any Award;
(vi) subject
to the provisions of Section 5(c) or 6(d), to extend at any time the period in which Stock Options or Stock Appreciation Rights,
respectively, may be exercised; and
(vii) at
any time to adopt, alter and repeal such rules, guidelines and practices for administration of the Plan and for its own acts and proceedings
as it shall deem advisable; to interpret the terms and provisions of the Plan and any Award (including related written instruments); to
make all determinations it deems advisable for the administration of the Plan; to decide all disputes arising in connection with the Plan;
and to otherwise supervise the administration of the Plan.
All decisions and interpretations
of the Administrator shall be binding on all persons, including the Company and Plan grantees.
(c) Delegation
of Authority to Grant Awards. Subject to applicable law, the Administrator, in its discretion, may delegate to a committee consisting
of one or more officers of the Company all or part of the Administrator’s authority and duties with respect to the granting of Awards
to individuals who are (i) not subject to the reporting and other provisions of Section 16 of the Exchange Act and (ii) not
members of the delegated committee. Any such delegation by the Administrator shall include a limitation as to the amount of Stock underlying
Awards that may be granted during the period of the delegation and shall contain guidelines as to the determination of the exercise price
and the vesting criteria. The Administrator may revoke or amend the terms of a delegation at any time, but such action shall not invalidate
any prior actions of the Administrator’s delegate or delegates that were consistent with the terms of the Plan.
(d) Award
Agreement. Awards under the Plan shall be evidenced by Award Agreements that set forth the terms, conditions and limitations for each
Award which may include, without limitation, the term of an Award and the provisions applicable in the event the Service Relationship
terminates.
(e) Indemnification.
Neither the Board nor the Administrator, nor any member of either or any delegate thereof, shall be liable for any act, omission, interpretation,
construction or determination made in good faith in connection with the Plan, and the members of the Board and the Administrator (and
any delegate thereof) shall be entitled in all cases to indemnification and reimbursement by the Company in respect of any claim, loss,
damage or expense (including, without limitation, reasonable attorneys’ fees) arising or resulting therefrom to the fullest extent
permitted by law and/or under the Company’s articles or bylaws or any directors’ and officers’ liability insurance coverage
which may be in effect from time to time and/or any indemnification agreement between such individual and the Company.
(f) Non-U.S.
Award Recipients. Notwithstanding any provision of the Plan to the contrary, in order to comply with the laws in other countries in
which the Company and its Affiliates operate or have employees or other individuals eligible for Awards, the Administrator, in its sole
discretion, shall have the power and authority to: (i) determine which Affiliates shall be covered by the Plan; (ii) determine
which individuals outside the United States are eligible to participate in the Plan; (iii) modify the terms and conditions of any
Award granted to individuals outside the United States to comply with applicable foreign laws; (iv) establish subplans and modify
exercise procedures and other terms and procedures, to the extent the Administrator determines such actions to be necessary or advisable
(and such subplans and/or modifications shall be incorporated into and made part of this Plan); provided, however, that no such subplans
and/or modifications shall increase the share limitations contained in Section 3(a) hereof; and (v) take any action, before
or after an Award is made, that the Administrator determines to be necessary or advisable to obtain approval or comply with any local
governmental regulatory exemptions or approvals. Notwithstanding the foregoing, the Administrator may not take any actions hereunder,
and no Awards shall be granted, that would violate the Exchange Act or any other applicable United States securities law, the Code, or
any other applicable United States governing statute or law.
SECTION 3. STOCK
ISSUABLE UNDER THE PLAN; MERGERS; SUBSTITUTION
(a) Stock
Issuable. The maximum number of shares of Stock reserved and available for issuance under the Plan shall be the sum of (i) 5,033,227
shares (the “Initial Limit”) and (ii) the number of shares reserved and remaining available for issuance under the Prior
Plan as of the Effective Date, plus on January 1, 2027 and on each January 1 thereafter, the number of shares of Stock reserved
and available for issuance under the Plan shall be cumulatively increased by 5% of the number of Outstanding Shares on the immediately
preceding December 31, or such lesser number of shares as approved by the Administrator, in all cases, subject to adjustment as provided
in this Section 3 (the “Annual Increase”). Subject to such overall limitation, the maximum aggregate number of shares
of Stock that may be issued in the form of Incentive Stock Options shall not exceed the Initial Limit, cumulatively increased on January 1,
2027 and on each January 1 thereafter by the lesser of the Annual Increase for such year or 5,033,227 shares of Stock, subject in
all cases to adjustment as provided in this Section 3. For purposes of this limitation, the shares of Stock underlying any awards
under the Plan and under the Prior Plan that are forfeited, canceled, held back upon exercise of an Option or settlement of an Award to
cover the exercise price or tax withholding, reacquired by the Company prior to vesting, satisfied without the issuance of Stock or otherwise
terminated (other than by exercise) shall be added back to the shares of Stock available for issuance under the Plan and, to the extent
permitted under Section 422 of the Code and the regulations promulgated thereunder, the shares of Stock that may be issued as Incentive
Stock Options. In the event the Company repurchases shares of Stock on the open market, such shares shall not be added to the shares of
Stock available for issuance under the Plan. Subject to such overall limitations, shares of Stock may be issued up to such maximum number
pursuant to any type or types of Award. The shares available for issuance under the Plan may be authorized but unissued shares of Stock
or shares of Stock reacquired by the Company. Awards that may be settled solely in cash shall not be counted against the share reserve,
nor shall they reduce the shares of Stock authorized for grant to a grantee in any calendar year.
(b) Maximum
Awards to Non-Employee Directors. Notwithstanding anything to the contrary in this Plan, the value of all Awards awarded under this
Plan and all other cash compensation paid by the Company to any Non-Employee Director for services as a Non-Employee Director in any calendar
year shall not exceed (i) $1,000,000 in the first calendar year an individual becomes a Non-Employee Director and (ii) $750,000
in any other calendar year. For purposes of this limitation, the value of any Award shall be its grant date fair value, as determined
in accordance with ASC Topic 718 or successor provision but excluding the impact of estimated forfeitures related to service-based vesting
provisions.
(c) Changes
in Stock. Subject to Section 3(d) hereof, if, as a result of any reorganization, recapitalization, reclassification, stock
dividend, extraordinary cash dividend, stock split, reverse stock split or other similar change in the Company’s capital stock,
the outstanding shares of Stock are increased or decreased or are exchanged for a different number or kind of shares or other securities
of the Company, or additional shares or new or different shares or other securities of the Company or other non-cash assets are distributed
with respect to such shares of Stock or other securities, or, if, as a result of any merger or consolidation, sale of all or substantially
all of the assets of the Company, the outstanding shares of Stock are converted into or exchanged for securities of the Company or any
successor entity (or a parent or subsidiary thereof), the Administrator shall make an appropriate or proportionate adjustment in (i) the
maximum number of shares reserved for issuance under the Plan, including the maximum number of shares that may be issued in the form of
Incentive Stock Options, (ii) the number and kind of shares or other securities subject to any then outstanding Awards under the
Plan, (iii) the repurchase price, if any, per share subject to each outstanding Restricted Stock Award and (iv) the exercise
price for each share subject to any then outstanding Stock Options and Stock Appreciation Rights under the Plan, without changing the
aggregate exercise price (i.e., the exercise price multiplied by the number of shares subject to Stock Options and Stock Appreciation
Rights) as to which such Stock Options and Stock Appreciation Rights remain exercisable. The Administrator shall also make equitable or
proportionate adjustments in the number of shares subject to outstanding Awards and the exercise price and the terms of outstanding Awards
to take into consideration cash dividends paid other than in the ordinary course or any other extraordinary corporate event. The adjustment
by the Administrator shall be final, binding and conclusive. No fractional shares of Stock shall be issued under the Plan resulting from
any such adjustment, but the Administrator in its discretion, may make a cash payment in lieu of fractional shares.
(d) Mergers
and Other Transactions. In the case of and subject to the consummation of a Sale Event, the Administrator, on such terms and conditions
as it deems appropriate, either by the terms of the Award or by action taken prior to the occurrence of such transaction or event and
without a grantee’s consent, is hereby authorized to take any one or more of the following actions: (i) to provide for the
assumption or continuation of Awards theretofore granted by the successor entity, or the substitution of such Awards with new Awards of
the successor entity or parent thereof, with appropriate adjustment as to the number and kind of shares and, if appropriate, the per share
exercise prices, as such parties shall agree; (ii) to provide that such Award shall vest and become exercisable, realizable, or payable,
or restrictions applicable to an Award will lapse, in whole or in part prior to or upon consummation of such Sale Event, and, to the extent
the Administrator determines, terminate upon or immediately prior to the effectiveness of such Sale Event and, to the extent applicable,
be exercisable as to all shares covered thereby, notwithstanding anything to the contrary in the Plan or the provisions of such Award;
(iii) to permit the exercise of Options and Stock Appreciation Rights (to the extent then vested and exercisable) for a limited period
of time prior to the consummation of the Sale Event following which, if not so exercised, such Award shall terminate as of the Sale Event;
(iv) to make or provide for a payment, in cash or in kind, to the grantees holding Options and Stock Appreciation Rights, in exchange
for the cancellation thereof, in an amount equal to the difference between (A) the Sale Price multiplied by the number of shares
of Stock subject to outstanding Options and Stock Appreciation Rights (to the extent then vested and exercisable at prices not in excess
of the Sale Price) and (B) the aggregate exercise price of all such outstanding Options and Stock Appreciation Rights (provided that,
in the case of an Option or Stock Appreciation Right with an exercise price equal to or greater than the Sale Price, such Option or Stock
Appreciation Right shall be cancelled for no consideration); (v) to make or provide for a payment, in cash or in kind, to the grantees
holding other Awards, in exchange for the cancellation thereof, in an amount equal to the Sale Price multiplied by the number of vested
shares of Stock under such Awards; (vi) with respect only to an Award (or portion thereof) that is unvested as of immediately prior
to the effective time of the Sale Event, the termination of the Award immediately prior to the Sale Event with such payment to the grantee
(including no payment) as the Administrator determines in its discretion; or (vii) any combination of the foregoing. In taking any
of the actions permitted under this subsection 3(d), the Administrator will not be obligated to treat all Awards, all Awards held by a
Participant, or all Awards of the same type, similarly.
(e) Substitute
Awards. Substitute Awards shall not reduce the shares of Stock authorized for grant under the Plan, nor shall shares subject to a
Substitute Award be added to the shares of Stock available for Awards under the Plan as provided in Section 3(a) above. Additionally,
in the event that a company acquired by the Company or any Affiliate or with which the Company or any Affiliate combines has shares available
under a pre-existing plan approved by stockholders and not adopted in contemplation of such acquisition or combination, the shares available
for grant pursuant to the terms of such pre-existing plan (as adjusted, to the extent appropriate, using the exchange ratio or other adjustment
or valuation ratio or formula used in such acquisition or combination to determine the consideration payable to the holders of common
stock of the entities party to such acquisition or combination) may be used for Awards under the Plan and shall not reduce the shares
authorized for grant under the Plan (and shares subject to such Awards shall not be added to the shares available for Awards under the
Plan as provided in Section 3(a) above); provided that Awards using such available shares shall not be made after the date awards
or grants could have been made under the terms of the pre-existing plan, absent the acquisition or combination, and shall only be made
to individuals who were not employees or directors of the Company or any of its Subsidiaries prior to such acquisition or combination.
SECTION 4. ELIGIBILITY
Grantees under the Plan will
be such employees, Non-Employee Directors or Consultants of the Company and its Affiliates as are selected from time to time by the Administrator
in its sole discretion; provided that Awards may not be granted to employees, Non-Employee Directors or Consultants who are providing
services only to any “parent” of the Company, as such term is defined in Rule 405 of the Act, unless (i) the stock
underlying the Awards is treated as “service recipient stock” under Section 409A or (ii) the Company has determined
that such Awards are exempt from or otherwise comply with Section 409A.
SECTION 5. STOCK
OPTIONS
(a) Award
of Stock Options. The Administrator may grant Stock Options under the Plan. Any Stock Option granted under the Plan shall be in such
form as the Administrator may from time to time approve.
Stock Options granted under
the Plan may be either Incentive Stock Options or Non-Qualified Stock Options. Incentive Stock Options may be granted only to employees
of the Company or any Subsidiary that is a “subsidiary corporation” within the meaning of Section 424(f) of the
Code. To the extent that any Option does not qualify as an Incentive Stock Option, it shall be deemed a Non-Qualified Stock Option.
Stock Options granted pursuant
to this Section 5 shall be subject to the following terms and conditions and shall contain such additional terms and conditions,
not inconsistent with the terms of the Plan, as the Administrator shall deem desirable. If the Administrator so determines, Stock Options
may be granted in lieu of cash compensation at the optionee’s election, subject to such terms and conditions as the Administrator
may establish.
(b) Exercise
Price. The exercise price per share for the Stock covered by a Stock Option granted pursuant to this Section 5 shall be determined
by the Administrator at the time of grant but shall not be less than 100 percent (100%) of the Fair Market Value on the date of grant.
In the case of an Incentive Stock Option that is granted to a Ten Percent Owner, the exercise price of such Incentive Stock Option shall
be not less than 110 percent (110%) of the Fair Market Value on the grant date. Notwithstanding the foregoing, Stock Options may be granted
with an exercise price per share that is less than 100 percent (100%) of the Fair Market Value on the date of grant (i) pursuant
to a transaction described in, and in a manner consistent with, Section 424(a) of the Code, (ii) to individuals who are
not subject to U.S. income tax on the date of grant or (iii) if the Stock Option is compliant with Section 409A.
(c) Option
Term. The term of each Stock Option shall be fixed by the Administrator, but no Stock Option shall be exercisable more than ten years
after the date the Stock Option is granted. In the case of an Incentive Stock Option that is granted to a Ten Percent Owner, the term
of such Stock Option shall be no more than five years from the date of grant.
(d) Exercisability;
Rights of a Stockholder. Stock Options shall become exercisable at such time or times, whether or not in installments, as shall be
determined by the Administrator at or after the grant date. The Administrator may at any time accelerate the exercisability of all or
any portion of any Stock Option. An optionee shall have the rights of a stockholder only as to shares acquired upon the exercise of a
Stock Option and not as to unexercised Stock Options.
(e) Method
of Exercise. Stock Options may be exercised in whole or in part, by giving written or electronic notice of exercise to the Company,
specifying the number of shares to be purchased. Payment of the purchase price may be made by one or more of the following methods except
to the extent otherwise provided in the Award Agreement:
(i) In
cash, by certified or bank check or other instrument acceptable to the Administrator;
(ii) Through
the delivery (or attestation to the ownership following such procedures as the Company may prescribe) of shares of Stock that are not
then subject to restrictions under any Company plan. Such surrendered shares shall be valued at Fair Market Value on the exercise date;
(iii) By
the optionee delivering to the Company a properly executed exercise notice together with irrevocable instructions to a broker to promptly
deliver to the Company cash or a check payable and acceptable to the Company for the purchase price; provided that in the event the optionee
chooses to pay the purchase price as so provided, the optionee and the broker shall comply with such procedures and enter into such agreements
of indemnity and other agreements as the Company shall prescribe as a condition of such payment procedure; or
(iv) With
respect to Stock Options that are not Incentive Stock Options, by a “net exercise” arrangement pursuant to which the Company
will reduce the number of shares of Stock issuable upon exercise by the largest whole number of shares with a Fair Market Value that does
not exceed the aggregate exercise price.
Payment instruments will be received subject to
collection. The transfer to the optionee on the records of the Company or of the transfer agent of the shares of Stock to be purchased
pursuant to the exercise of a Stock Option will be contingent upon receipt from the optionee (or a purchaser acting in his stead in accordance
with the provisions of the Stock Option) by the Company of the full purchase price for such shares and the fulfillment of any other requirements
contained in the Award Agreement or applicable provisions of laws (including the satisfaction of any taxes that the Company or an Affiliate
is obligated to withhold with respect to the optionee). In the event an optionee chooses to pay the purchase price by previously-owned
shares of Stock through the attestation method, the number of shares of Stock transferred to the optionee upon the exercise of the Stock
Option shall be net of the number of attested shares. In the event that the Company establishes, for itself or using the services of a
third party, an automated system for the exercise of Stock Options, such as a system using an internet website or interactive voice response,
then the paperless exercise of Stock Options may be permitted through the use of such an automated system.
(f) Annual
Limit on Incentive Stock Options. To the extent required for “incentive stock option” treatment under Section 422
of the Code, the aggregate Fair Market Value (determined as of the time of grant) of the shares of Stock with respect to which Incentive
Stock Options granted under this Plan and any other plan of the Company or its parent and subsidiary corporations become exercisable for
the first time by an optionee during any calendar year shall not exceed $100,000. To the extent that any Stock Option exceeds this limit,
it shall constitute a Non-Qualified Stock Option.
SECTION 6. STOCK
APPRECIATION RIGHTS
(a) Award
of Stock Appreciation Rights. The Administrator may grant Stock Appreciation Rights under the Plan. A Stock Appreciation Right is
an Award entitling the recipient to receive shares of Stock (or cash, to the extent explicitly provided for in the applicable Award Agreement)
having a value equal to the excess of the Fair Market Value of a share of Stock on the date of exercise over the exercise price of the
Stock Appreciation Right multiplied by the number of shares of Stock with respect to which the Stock Appreciation Right shall have been
exercised.
(b) Exercise
Price of Stock Appreciation Rights. The exercise price of a Stock Appreciation Right shall not be less than 100 percent (100%) of
the Fair Market Value of the Stock on the date of grant. Notwithstanding the foregoing, Stock Appreciation Rights may be granted with
an exercise price per share that is less than 100 percent (100%) of the Fair Market Value on the date of grant (i) pursuant to a
transaction described in, and in a manner consistent with, Section 424(a) of the Code, (ii) to individuals who are not
subject to U.S. income tax on the date of grant or (iii) if the Stock Appreciation Rights is compliant with Section 409A.
(c) Grant
and Exercise of Stock Appreciation Rights. Stock Appreciation Rights may be granted by the Administrator independently of any Stock
Option granted pursuant to Section 5 of the Plan.
(d) Terms
and Conditions of Stock Appreciation Rights. Stock Appreciation Rights shall be subject to such terms and conditions as shall be determined
on the date of grant by the Administrator. The term of a Stock Appreciation Right may not exceed ten years. The terms and conditions of
each such Award shall be determined by the Administrator, and such terms and conditions may differ among individual Awards and grantees.
SECTION 7. RESTRICTED
STOCK AWARDS
(a) Nature
of Restricted Stock Awards. The Administrator may grant Restricted Stock Awards under the Plan. A Restricted Stock Award is any Award
of Restricted Shares subject to such restrictions and conditions as the Administrator may determine at the time of grant. Conditions may
be based on continuing employment (or other Service Relationship) and/or achievement of pre-established performance goals and objectives.
(b) Rights
as a Stockholder. Upon the grant of the Restricted Stock Award and payment of any applicable purchase price, a grantee shall have
the rights of a stockholder with respect to the voting of the Restricted Shares and receipt of dividends; provided that if the lapse of
restrictions with respect to the Restricted Stock Award is tied to the attainment of vesting conditions, any dividends paid by the Company
shall accrue and shall not be paid to the grantee until and to the extent the vesting conditions are met with respect to the Restricted
Stock Award. Unless the Administrator shall otherwise determine, (i) uncertificated Restricted Shares shall be accompanied by a notation
on the records of the Company or the transfer agent to the effect that they are subject to forfeiture until such Restricted Shares are
vested as provided in Section 7(d) below, and (ii) certificated Restricted Shares shall remain in the possession of the
Company until such Restricted Shares are vested as provided in Section 7(d) below, and the grantee shall be required, as a condition
of the grant, to deliver to the Company such instruments of transfer as the Administrator may prescribe.
(c) Restrictions.
Restricted Shares may not be sold, assigned, transferred, pledged or otherwise encumbered or disposed of except as specifically provided
herein or in the Restricted Stock Award Agreement. Except as may otherwise be provided by the Administrator either in the Award Agreement
or, subject to Section 16 below, in writing after the Award is issued, if a grantee’s employment (or other Service Relationship)
with the Company and its Affiliates terminates for any reason, any Restricted Shares that have not vested at the time of termination shall
automatically and without any requirement of notice to such grantee from or other action by or on behalf of, the Company be deemed to
have been reacquired by the Company at its original purchase price (if any) from such grantee or such grantee’s legal representative
simultaneously with such termination of employment (or other Service Relationship), and thereafter shall cease to represent any ownership
of the Company by the grantee or rights of the grantee as a stockholder. Following such deemed reacquisition of Restricted Shares that
are represented by physical certificates, a grantee shall surrender such certificates to the Company upon request without consideration.
(d) Vesting
of Restricted Shares. The Administrator at the time of grant shall specify the date or dates and/or the attainment of pre-established
performance goals, objectives and other conditions on which the non-transferability of the Restricted Shares and the Company’s right
of repurchase or forfeiture shall lapse. Subsequent to such date or dates and/or the attainment of such pre-established performance goals,
objectives and other conditions, the shares on which all restrictions have lapsed shall no longer be Restricted Shares and shall be deemed
“vested.”
SECTION 8. RESTRICTED
STOCK UNITS
(a) Nature
of Restricted Stock Units. The Administrator may grant Restricted Stock Units under the Plan. A Restricted Stock Unit is an Award
of stock units that may be settled in shares of Stock (or cash, to the extent explicitly provided for in the Award Agreement) upon the
satisfaction of such restrictions and conditions at the time of grant. Conditions may be based on continuing employment (or other Service
Relationship) and/or achievement of pre-established performance goals and objectives. The terms and conditions of each such Award shall
be determined by the Administrator, and such terms and conditions may differ among individual Awards and grantees. Restricted Stock Units
with deferred settlement dates are subject to Section 409A and shall contain such additional terms and conditions as the Administrator
shall determine in its sole discretion in order to comply with the requirements of Section 409A.
(b) Election
to Receive Restricted Stock Units in Lieu of Compensation. The Administrator may, in its sole discretion, permit a grantee to elect
to receive a portion of future cash compensation otherwise due to such grantee in the form of an award of Restricted Stock Units. Any
such election shall be made in writing and shall be delivered to the Company no later than the date specified by the Administrator and
in accordance with Section 409A and such other rules and procedures established by the Administrator. Any such future cash compensation
that the grantee elects to defer shall be converted to a fixed number of Restricted Stock Units based on the Fair Market Value of Stock
on the date the compensation would otherwise have been paid to the grantee if such payment had not been deferred as provided herein. The
Administrator shall have the sole right to determine whether and under what circumstances to permit such elections and to impose such
limitations and other terms and conditions thereon as the Administrator deems appropriate. Any Restricted Stock Units that are elected
to be received in lieu of cash compensation shall be fully vested, unless otherwise provided in the Award Agreement.
(c) Rights
as a Stockholder. A grantee shall have the rights as a stockholder only as to shares of Stock acquired by the grantee upon settlement
of Restricted Stock Units; provided, however, that the grantee may be credited with Dividend Equivalent Rights with respect to the stock
units underlying his or her Restricted Stock Units, subject to the provisions of Section 11 and such terms and conditions as the
Administrator may determine.
(d) Termination.
Except as may otherwise be provided by the Administrator either in the Award Agreement or, subject to Section 16 below, in writing
after the Award is issued, a grantee’s right in all Restricted Stock Units that have not vested shall automatically terminate upon
the grantee’s termination of employment (or cessation of Service Relationship) with the Company and its Affiliates for any reason.
SECTION 9. UNRESTRICTED
STOCK AWARDS
Grant
or Sale of Unrestricted Stock. The Administrator may grant (or sell at par value or such higher purchase price determined by
the Administrator) an Unrestricted Stock Award under the Plan. An Unrestricted Stock Award is an Award pursuant to which the grantee may
receive shares of Stock free of any restrictions under the Plan. Unrestricted Stock Awards may be granted in respect of past services
or other valid consideration, or in lieu of cash compensation due to such grantee.
SECTION 10. CASH-BASED
AWARDS
Grant
of Cash-Based Awards. The Administrator may grant Cash-Based Awards under the Plan. A Cash-Based Award is an Award that entitles
the grantee to a payment in cash upon the attainment of specified performance goals, including continued employment (or other Service
Relationship). The Administrator shall determine the maximum duration of the Cash-Based Award, the amount of cash to which the Cash-Based
Award pertains, the conditions upon which the Cash-Based Award shall become vested or payable, and such other provisions as the Administrator
shall determine. Each Cash-Based Award shall specify a cash-denominated payment amount, formula or payment ranges as determined by the
Administrator. Payment, if any, with respect to a Cash-Based Award shall be made in accordance with the terms of the Award and may be
made in cash.
SECTION 11. DIVIDEND
EQUIVALENT RIGHTS
(a) Dividend
Equivalent Rights. The Administrator may grant Dividend Equivalent Rights under the Plan. A Dividend Equivalent Right is an Award
entitling the grantee to receive credits based on cash dividends that would have been paid on the shares of Stock specified in the Dividend
Equivalent Right (or other Award to which it relates) if such shares had been issued to the grantee. A Dividend Equivalent Right may be
granted hereunder to any grantee as a component of an award of Restricted Stock Units or as a freestanding award. The terms and conditions
of Dividend Equivalent Rights shall be specified in the Award Agreement. Dividend equivalents credited to the holder of a Dividend Equivalent
Right may be paid currently or may be deemed to be reinvested in additional shares of Stock, which may thereafter accrue additional equivalents.
Any such reinvestment shall be at Fair Market Value on the date of reinvestment or such other price as may then apply under a dividend
reinvestment plan sponsored by the Company, if any. Dividend Equivalent Rights may be settled in cash or shares of Stock or a combination
thereof, in a single installment or installments. A Dividend Equivalent Right granted as a component of an Award of Restricted Stock Units
shall provide that such Dividend Equivalent Right shall be settled only upon settlement or payment of, or lapse of restrictions on, such
other Award, and that such Dividend Equivalent Right shall expire or be forfeited or annulled under the same conditions as such other
Award.
(b) Termination.
Except as may otherwise be provided by the Administrator either in the Award Agreement or, subject to Section 16 below, in writing
after the Award is issued, a grantee’s rights in all Dividend Equivalent Rights shall automatically terminate upon the grantee’s
termination of employment (or cessation of Service Relationship) with the Company and its Affiliates for any reason.
SECTION 12. Transferability
of Awards
(a) Transferability.
Except as provided in Section 12(b) below or otherwise determined by the Administrator, during a grantee’s lifetime, his
or her Awards shall be exercisable only by the grantee, or by the grantee’s legal representative or guardian in the event of the
grantee’s incapacity. No Awards shall be sold, assigned, transferred or otherwise encumbered or disposed of by a grantee other than
by will or by the laws of descent and distribution or pursuant to a domestic relations order. No Awards shall be subject, in whole or
in part, to attachment, execution, or levy of any kind, and any purported transfer in violation hereof shall be null and void.
(b) Administrator
Action. Notwithstanding Section 12(a), the Administrator, in its discretion, may provide either in the Award Agreement regarding
a given Award or by subsequent written approval that the grantee (who is an employee or Non-Employee Director) may transfer his or her
Non-Qualified Stock Options to his or her immediate family members, to trusts for the benefit of such family members, or to partnerships
in which such family members are the only partners, provided that the transferee agrees in writing with the Company to be bound by all
of the terms and conditions of this Plan and the applicable Award Agreement. In no event may an Award be transferred by a grantee for
value.
(c) Family
Member. For purposes of Section 12(b), “family member” shall mean a grantee’s child, stepchild, grandchild,
parent, stepparent, grandparent, spouse, former spouse, sibling, niece, nephew, mother-in-law, father-in-law, son-in-law, daughter-in-law,
brother-in-law, or sister-in-law, including adoptive relationships, any person sharing the grantee’s household (other than a tenant
of the grantee), a trust in which these persons (or the grantee) have more than 50 percent (50%) of the beneficial interest, a foundation
in which these persons (or the grantee) control the management of assets, and any other entity in which these persons (or the grantee)
own more than 50 percent (50%) of the voting interests.
(d) Designation
of Beneficiary. To the extent permitted by the Company and valid under applicable law, each grantee to whom an Award has been made
under the Plan may designate a beneficiary or beneficiaries to exercise any Award or receive any payment under any Award payable on or
after the grantee’s death. Any such designation shall be on a form provided for that purpose by the Administrator and shall not
be effective until received by the Administrator. If no beneficiary has been designated by a deceased grantee, or if the designated beneficiaries
have predeceased the grantee, the beneficiary shall be the grantee’s estate or legal heirs.
SECTION 13. TAX
WITHHOLDING
(a) Payment
by Grantee. Each grantee shall, no later than the date as of which the value of an Award or of any Stock or other amounts received
thereunder first becomes includable in the gross income of the grantee for tax purposes, pay to the Company or any applicable Affiliate,
or make arrangements satisfactory to the Administrator regarding payment of, any U.S. and non-U.S. federal, state, or local taxes of any
kind required by law to be withheld by the Company or any applicable Affiliate with respect to such income. The Company and its Affiliates
shall, to the extent permitted by law, have the right to deduct any such taxes from any payment of any kind otherwise due to the grantee
or to satisfy any applicable withholding obligations by any other method of withholding that the Company and its Affiliates deem appropriate.
The Company’s obligation to deliver evidence of book entry (or stock certificates) to any grantee is subject to and conditioned
on tax withholding obligations being satisfied by the grantee.
(b) Payment
in Stock. The Administrator may cause any tax withholding obligation of the Company or any applicable Affiliate to be satisfied, in
whole or in part, by the Company withholding from shares of Stock to be issued pursuant to any Award a number of shares with an aggregate
Fair Market Value (as of the date the withholding is effected) that would satisfy the withholding amount due; provided, however, that
the amount withheld does not exceed the maximum statutory rate or such lesser amount as is necessary to avoid liability accounting treatment.
For purposes of share withholding, the Fair Market Value of withheld shares shall be determined in the same manner as the value of Stock
includible in income of the grantees. The Administrator may also require any tax withholding obligation of the Company or any applicable
Affiliate to be satisfied, in whole or in part, by an arrangement whereby a certain number of shares of Stock issued pursuant to any Award
are immediately sold and proceeds from such sale are remitted to the Company or any applicable Affiliate in an amount that would satisfy
the withholding amount due.
SECTION 14. Section 409A
awards
Awards are intended to be
exempt from Section 409A to the greatest extent possible and to otherwise comply with Section 409A. The Plan and all Awards
shall be interpreted in accordance with such intent. To the extent that any Award is determined to constitute “nonqualified deferred
compensation” within the meaning of Section 409A (a “409A Award”), the Award shall be subject to such additional
rules and requirements as specified by the Administrator from time to time in order to comply with Section 409A. In this regard,
if any amount under a 409A Award is payable upon a “separation from service” (within the meaning of Section 409A) to
a grantee who is then considered a “specified employee” (within the meaning of Section 409A), then no such payment shall
be made prior to the date that is the earlier of (i) six months and one day after the grantee’s separation from service, or
(ii) the grantee’s death, but only to the extent such delay is necessary to prevent such payment from being subject to interest,
penalties and/or additional tax imposed pursuant to Section 409A. Further, the settlement of any 409A Award may not be accelerated
except to the extent permitted by Section 409A. The Company makes no representation that any or all of the payments or benefits described
in the Plan will be exempt from or comply with Section 409A of the Code and makes no undertaking to preclude Section 409A of
the Code from applying to any such payment. The grantee shall be solely responsible for the payment of any taxes and penalties incurred
under Section 409A.
SECTION 15. TERMINATION
OF SERVICE RELATIONSHIP, TRANSFER, LEAVE OF ABSENCE, ETC.
(a) Termination
of Service Relationship. If the grantee’s Service Relationship is with an Affiliate and such Affiliate ceases to be an Affiliate,
the grantee shall be deemed to have terminated his or her Service Relationship for purposes of the Plan.
(b) For
purposes of the Plan, the following events shall not be deemed a termination of a Service Relationship:
(i) a
transfer of a Service Relationship from the Company to an Affiliate or from an Affiliate to the Company, or from one Affiliate to another;
or
(ii) an
approved leave of absence for military service or sickness, or for any other purpose approved by the Company, if the employee’s
right to re-employment is guaranteed either by a statute or by contract or under the policy pursuant to which the leave of absence was
granted or if the Administrator otherwise so provides in writing.
SECTION 16. AMENDMENTS
AND TERMINATION
The Board may, at any time,
amend or discontinue the Plan and the Administrator may, at any time, amend or cancel any outstanding Award for the purpose of satisfying
changes in law or for any other lawful purpose, but no such action shall materially and adversely affect rights under any outstanding
Award without the holder’s consent. The Board is specifically authorized, without the consent of any holder and without the consent
of the Company’s stockholders, to exercise its discretion to reduce the exercise price of outstanding Stock Options or Stock Appreciation
Rights, effect the repricing of such Awards through cancellation and re-grants or cancel such Awards in exchange for cash or other Awards.
To the extent required under the rules of any securities exchange or market system on which the Stock is listed, to the extent determined
by the Administrator to be required by the Code to ensure that Incentive Stock Options granted under the Plan are qualified under Section 422
of the Code, Plan amendments shall be subject to approval by Company stockholders. Nothing in this Section 16 shall limit the Administrator’s
authority to take any action permitted pursuant to Section 3(c) or 3(d).
SECTION 17. STATUS
OF PLAN
With respect to the portion
of any Award that has not been exercised and any payments in cash, Stock or other consideration not received by a grantee, a grantee shall
have no rights greater than those of a general creditor of the Company unless the Administrator shall otherwise expressly determine in
connection with any Award or Awards. In its sole discretion, the Administrator may authorize the creation of trusts or other arrangements
to meet the Company’s obligations to deliver Stock or make payments with respect to Awards hereunder, provided that the existence
of such trusts or other arrangements is consistent with the foregoing sentence.
SECTION 18. GENERAL
PROVISIONS
(a) No
Distribution. The Administrator may require each person acquiring Stock pursuant to an Award to represent to and agree with the Company
in writing that such person is acquiring the shares without a view to distribution thereof.
(b) Issuance
of Stock. To the extent certificated, stock certificates to grantees under this Plan shall be deemed delivered for all purposes when
the Company or a stock transfer agent of the Company shall have mailed such certificates in the United States mail, addressed to the grantee,
at the grantee’s last known address on file with the Company. Uncertificated Stock shall be deemed delivered for all purposes when
the Company or a Stock transfer agent of the Company shall have given to the grantee by electronic mail (with proof of receipt) or by
United States mail, addressed to the grantee, at the grantee’s last known address on file with the Company, notice of issuance and
recorded the issuance in its records (which may include electronic “book entry” records). Notwithstanding anything herein
to the contrary, the Company shall not be required to issue or deliver any evidence of book entry or certificates evidencing shares of
Stock pursuant to the exercise or settlement of any Award, unless and until the Administrator has determined, with advice of counsel (to
the extent the Administrator deems such advice necessary or advisable), that the issuance and delivery is in compliance with all applicable
laws, regulations of governmental authorities and, if applicable, the requirements of any exchange on which the shares of Stock are listed,
quoted or traded. Any Stock issued pursuant to the Plan shall be subject to any stop-transfer orders and other restrictions as the Administrator
deems necessary or advisable to comply with federal, state or foreign jurisdiction, securities or other laws, rules and quotation
system on which the Stock is listed, quoted or traded. The Administrator may place legends on any Stock certificate or notations on any
book entry to reference restrictions applicable to the Stock. In addition to the terms and conditions provided herein, the Administrator
may require that an individual make such reasonable covenants, agreements, and representations as the Administrator, in its discretion,
deems necessary or advisable in order to comply with any such laws, regulations, or requirements. The Administrator shall have the right
to require any individual to comply with any timing or other restrictions with respect to the settlement or exercise of any Award, including
a window-period limitation, as may be imposed in the discretion of the Administrator.
(c) Stockholder
Rights. Until Stock is deemed delivered in accordance with Section 18(b), no right to vote or receive dividends or any other
rights of a stockholder will exist with respect to shares of Stock to be issued in connection with an Award, notwithstanding the exercise
of a Stock Option or any other action by the grantee with respect to an Award.
(d) Other
Incentive Arrangements; No Rights to Continued Service Relationship. Nothing contained in this Plan shall prevent the Board from adopting
other or additional incentive arrangements, including trusts, and such arrangements may be either generally applicable or applicable only
in specific cases. The adoption of this Plan and the grant of Awards do not confer upon any grantee any right to continued employment
or other Service Relationship with the Company or any Affiliate.
(e) Trading
Policy Restrictions. Option exercises and other Awards under the Plan shall be subject to the Company’s insider trading policies
and procedures, as in effect from time to time.
(f) Clawback
Policy. Awards under the Plan shall be subject to the Company’s clawback policy, as in effect from time to time. In addition,
the Administrator may impose such other clawback, recovery, or recoupment provisions in an Award Agreement as the Administrator determines
necessary or appropriate, including, but not limited to, a reacquisition right in respect of previously acquired shares of Stock or other
cash or property upon the occurrence of a termination for “cause” under any agreement with the Company or an Affiliate thereof.
No recovery of compensation under such a clawback policy will be an event giving rise to a right to resign for “good reason”
or “constructive termination” (or similar term) under any agreement with the Company or an Affiliate thereof.
SECTION 19. EFFECTIVE
DATE OF PLAN
This Plan shall become effective
upon the date immediately preceding the Registration Date subject to prior stockholder approval in accordance with applicable state law,
the Company’s bylaws and articles of incorporation, and applicable stock exchange rules. No grants of Awards may be made hereunder
after the tenth anniversary of the Effective Date and no grants of Incentive Stock Options may be made hereunder after the tenth anniversary
of the date the Plan is approved by the Board.
SECTION 20. GOVERNING
LAW
This Plan and all Awards and
actions taken thereunder shall be governed by, and construed in accordance with, the General Corporation Law of the State of Delaware
as to matters within the scope thereof, and as to all other matters shall be governed by and construed in accordance with the internal
laws of the State of Delaware, applied without regard to conflict of law principles.
DATE APPROVED BY BOARD OF DIRECTORS: September 19, 2026
DATE APPROVED BY STOCKHOLDERS: October 8, 2026
INCENTIVE STOCK
OPTION AGREEMENT
UNDER THE CITY THERAPEUTICS, INC.
2026 STOCK OPTION AND INCENTIVE PLAN
| Name of Optionee: |
____________________________________ |
| No. of Option Shares: |
___________________ |
| Option Exercise Price per Share: |
$__________________ |
[FMV on Grant Date (110% of FMV if
a 10% owner)]
| Grant Date: |
__________________ |
| Expiration Date: |
__________________ |
[up to 10 years (5 if a 10% owner)]
Pursuant
to the City Therapeutics, Inc. 2026 Stock Option and Incentive Plan as amended through the date hereof (the “Plan”),
City Therapeutics, Inc. (the “Company”) hereby grants to the Optionee named above an option (the “Stock Option”)
to purchase on or prior to the Expiration Date specified above all or part of the number of shares of Common Stock, par value $0.0001
per share (the “Stock”), of the Company specified above at the Option Exercise Price per Share specified above subject to
the terms and conditions set forth herein and in the Plan.
1. Exercisability
Schedule. No portion of this Stock Option may be exercised until such portion shall have become exercisable. Except as set forth
below, and subject to the discretion of the Administrator (as defined in Section 2 of the Plan) to accelerate the exercisability
schedule hereunder, this Stock Option shall be exercisable as follows: ___________________, so long as the Optionee continues to have
a Service Relationship (as defined in the Plan) with the Company or a Subsidiary on such dates.
Once
exercisable, this Stock Option shall continue to be exercisable at any time or times prior to the close of business on the Expiration
Date, subject to the provisions hereof and of the Plan.
2. Manner
of Exercise.
(a) The
Optionee may exercise this Stock Option only in the following manner: from time to time on or prior to the Expiration Date of this Stock
Option, the Optionee may give written notice to the Administrator of his or her election to purchase some or all of the Option Shares
purchasable at the time of such notice. This notice shall specify the number of Option Shares to be purchased.
Payment
of the purchase price for the Option Shares may be made by one or more of the following methods: (i) in cash, by certified or bank
check or other instrument acceptable to the Administrator; (ii) through the delivery (or attestation to the ownership) of shares
of Stock that have been purchased by the Optionee on the open market or that are beneficially owned by the Optionee and are not then
subject to any restrictions under any Company plan and that otherwise satisfy any holding periods as may be required by the Administrator;
(iii) by the Optionee delivering to the Company a properly executed exercise notice together with irrevocable instructions to a
broker to promptly deliver to the Company cash or a check payable and acceptable to the Company to pay the option purchase price, provided
that in the event the Optionee chooses to pay the option purchase price as so provided, the Optionee and the broker shall comply with
such procedures and enter into such agreements of indemnity and other agreements as the Administrator shall prescribe as a condition
of such payment procedure; or (iv) a combination of (i), (ii) and (iii) above. Payment instruments will be received subject
to collection.
The
transfer to the Optionee on the records of the Company or of the transfer agent of the Option Shares will be contingent upon (i) the
Company’s receipt from the Optionee of the full purchase price for the Option Shares, as set forth above, (ii) the fulfillment
of any other requirements contained herein or in the Plan or in any other agreement or provision of laws and (iii) the receipt by
the Company of any agreement, statement or other evidence that the Company may require to satisfy itself that the issuance of Stock to
be purchased pursuant to the exercise of Stock Options under the Plan and any subsequent resale of the shares of Stock will be in compliance
with applicable laws and regulations. In the event the Optionee chooses to pay the purchase price by previously-owned shares of Stock
through the attestation method, the number of shares of Stock transferred to the Optionee upon the exercise of the Stock Option shall
be net of the shares of Stock attested to.
(b) The
shares of Stock purchased upon exercise of this Stock Option shall be transferred to the Optionee on the records of the Company or of
the transfer agent upon compliance to the satisfaction of the Administrator with all requirements under applicable laws or regulations
in connection with such transfer and with the requirements hereof and of the Plan. The determination of the Administrator as to such
compliance shall be final and binding on the Optionee. The Optionee shall not be deemed to be the holder of, or to have any of the rights
of a holder with respect to, any shares of Stock subject to this Stock Option unless and until this Stock Option shall have been exercised
pursuant to the terms hereof, the Company or the transfer agent shall have transferred the shares to the Optionee, and the Optionee’s
name shall have been entered as the stockholder of record on the books of the Company. Thereupon, the Optionee shall have full voting,
dividend and other ownership rights with respect to such shares of Stock.
(c) Notwithstanding
any other provision hereof or of the Plan, no portion of this Stock Option shall be exercisable after the Expiration Date hereof.
3. Termination
of Service Relationship. If the Optionee’s Service Relationship with the Company or a Subsidiary (as defined in the Plan) is
terminated, the period within which to exercise the Stock Option may be subject to earlier termination as set forth below.
(a) Termination
Due to Death. If the Optionee’s Service Relationship with the Company or a Subsidiary terminates by reason of the Optionee’s
death, any portion of this Stock Option outstanding on such date, to the extent exercisable on the date of death, may thereafter be exercised
by the Optionee’s legal representative or legatee for a period of twelve (12) months from the date of death or until the Expiration
Date, if earlier. Any portion of this Stock Option that is not exercisable on the date of death shall terminate immediately and be of
no further force or effect.
(b) Termination
Due to Disability. If the Optionee’s Service Relationship with the Company or a Subsidiary terminates by reason of the Optionee’s
disability (as determined by the Administrator), any portion of this Stock Option outstanding on such date, to the extent exercisable
on the date of such disability, may thereafter be exercised by the Optionee for a period of twelve (12) months from the date the Optionee’s
Service Relationship is terminated by reason of the Optionee’s disability or until the Expiration Date, if earlier. Any portion
of this Stock Option that is not exercisable on the date of the termination of the Optionee’s Service Relationship by reason of
the Optionee’s disability shall terminate immediately and be of no further force or effect.
(c) Termination
for Cause. If the Optionee’s Service Relationship with the Company or a Subsidiary terminates for Cause, any portion of this
Stock Option outstanding on such date shall terminate immediately and be of no further force and effect. For purposes hereof, “Cause”
shall mean, unless otherwise provided in an employment or service agreement between the Company and the Optionee, a determination by
the Administrator that the Optionee shall be dismissed as a result of (i) any material breach by the Optionee of any agreement between
the Optionee and the Company; (ii) the conviction of, indictment for or plea of nolo contendere by the Optionee to a felony or a
crime involving moral turpitude; or (iii) any material misconduct or willful and deliberate non-performance (other than by reason
of disability) by the Optionee of the Optionee’s duties to the Company.
(d) Other
Termination. If the Optionee’s Service Relationship with the Company or a Subsidiary terminates for any reason other than the
Optionee’s death, the Optionee’s disability, or Cause, and unless otherwise determined by the Administrator, any portion
of this Stock Option outstanding on such date may be exercised, to the extent exercisable on the date of termination, for a period of
ninety (90) days from the date of termination or until the Expiration Date, if earlier. Any portion of this Stock Option that is not
exercisable on the date of termination shall terminate immediately and be of no further force or effect.
The
Administrator’s determination of the reason for termination of the Optionee’s Service Relationship with the Company or a
Subsidiary shall be conclusive and binding on the Optionee and his or her representatives or legatees.
4. Incorporation
of Plan. Notwithstanding anything herein to the contrary, this Stock Option shall be subject to and governed by all the terms and
conditions of the Plan, including the powers of the Administrator set forth in Section 2(b) of the Plan. Capitalized terms
in this Agreement shall have the meaning specified in the Plan, unless a different meaning is specified herein.
5. Transferability.
This Agreement is personal to the Optionee, is non-assignable and is not transferable in any manner, by operation of law or otherwise,
other than by will or the laws of descent and distribution. This Stock Option is exercisable, during the Optionee’s lifetime, only
by the Optionee, and thereafter, only by the Optionee’s legal representative or legatee.
6. Status
of the Stock Option. This Stock Option is intended to qualify as an “incentive stock option” under Section 422 of
the Internal Revenue Code of 1986, as amended (the “Code”), but the Company does not represent or warrant that this Stock
Option qualifies as such. The Optionee should consult with his or her own tax advisors regarding the tax effects of this Stock Option
and the requirements necessary to obtain favorable income tax treatment under Section 422 of the Code, including, but not limited
to, holding period requirements and that this Stock Option must be exercised within three (3) months after termination of
employment as an employee (or twelve (12) months in the case of death or disability) to qualify as an “incentive stock option”.
To the extent any portion of this Stock Option does not so qualify as an “incentive stock option,” such portion shall be
deemed to be a non-qualified stock option. If the Optionee intends to dispose or does dispose (whether by sale, gift, transfer or otherwise)
of any Option Shares within the one-year period beginning on the date after the transfer of such shares to him or her, or within the
two-year period beginning on the day after the grant of this Stock Option, he or she will so notify the Company within 30 days after
such disposition.
7. Tax
Withholding. The Optionee shall, not later than the date as of which the exercise of this Stock Option becomes a taxable event for
Federal income tax purposes, pay to the Company or make arrangements satisfactory to the Administrator for payment of any Federal, state
and local taxes required by law to be withheld on account of such taxable event. The Company shall have the authority to cause the required
tax withholding obligation to be satisfied, in whole or in part, by (i) withholding from shares of Stock to be issued to the Optionee
a number of shares of Stock with an aggregate Fair Market Value that would satisfy the withholding amount due; or (ii) causing its
transfer agent to sell from the number of shares of Stock to be issued to the Optionee, the number of shares of Stock necessary to satisfy
the Federal, state and local taxes required by law to be withheld from the Optionee on account of such transfer.
8. No
Obligation to Continue Service Relationship. Neither the Company nor any Subsidiary is obligated by or as a result of the Plan or
this Agreement to continue the Optionee in a Service Relationship with the Company or any Subsidiary and neither the Plan nor this Agreement
shall interfere in any way with the right of the Company or any Subsidiary to terminate the Optionee’s Service Relationship with
the Company or a Subsidiary at any time.
9. Integration.
This Agreement constitutes the entire agreement between the parties with respect to this Stock Option and supersedes all prior agreements
and discussions between the parties concerning such subject matter.
10. Data
Privacy Consent. In order to administer the Plan and this Agreement and to implement or structure future equity grants, the Company,
its subsidiaries and affiliates and certain agents thereof (together, the “Relevant Companies”) may process any and all personal
or professional data, including but not limited to Social Security or other identification number, home address and telephone number,
date of birth and other information that is necessary or desirable for the administration of the Plan and/or this Agreement (the “Relevant
Information”). By entering into this Agreement, the Optionee (i) authorizes the Company to collect, process, register and
transfer to the Relevant Companies all Relevant Information; (ii) waives any privacy rights the Optionee may have with respect to
the Relevant Information; (iii) authorizes the Relevant Companies to store and transmit such information in electronic form; and
(iv) authorizes the transfer of the Relevant Information to any jurisdiction in which the Relevant Companies consider appropriate.
The Optionee shall have access to, and the right to change, the Relevant Information. Relevant Information will only be used in accordance
with applicable law.
11. Notices.
Notices hereunder shall be mailed or delivered to the Company at its principal place of business and shall be mailed or delivered to
the Optionee at the address on file with the Company or, in either case, at such other address as one party may subsequently furnish
to the other party in writing.
12. [Clawback
Acknowledgement. The Optionee acknowledges that the Optionee may become subject to the City Therapeutics, Inc. Compensation
Recovery Policy adopted pursuant to Rule 10D-1 promulgated under the Exchange Act and Nasdaq Rule 5608, or any successor rule (the
“Clawback Policy”). The Optionee understands that if the Optionee is or becomes subject to the Clawback Policy, the Company
and/or the Board shall be entitled to recover all Erroneously Awarded Compensation (as defined in the Clawback Policy) from the Optionee
pursuant to such means as the Company and/or the Board may elect. The Optionee agrees that the Optionee shall take all required action
to enable such recovery. The Optionee understands that such recovery may be sought and occur after the Optionee’s employment or
service with the Company terminates. The Optionee further agrees that the Optionee is not entitled to indemnification for any Erroneously
Awarded Compensation or for any claim or losses arising out of or in any way related to Erroneously Awarded Compensation recovered pursuant
to the Clawback Policy and, to the extent any agreement or organizational document purports to provide otherwise, the Optionee hereby
irrevocably agrees to forego such indemnification. The Optionee acknowledges and agrees that the Optionee has received and has had an
opportunity to review the Clawback Policy. Any action by the Company to recover Erroneously Awarded Compensation under the Clawback Policy
from the Optionee shall not, whether alone or in combination with any other action, event or condition, be deemed (i) an event giving
rise to a right to resign for Good Reason (as defined in the City Therapeutics, Inc. Executive Severance Plan) or serve as a basis
for a claim of constructive termination under any benefits or compensation arrangement applicable to the Optionee, or (ii) to constitute
a breach of a contract or other arrangement to which the Optionee is a party. This Section 12 is a material term of this Agreement.]1
1
Note to Draft: For Section 16 officers only.
| |
City Therapeutics, Inc. |
| |
|
|
| |
By: |
|
| |
|
Title: |
The foregoing Agreement
is hereby accepted and the terms and conditions thereof hereby agreed to by the undersigned. Electronic acceptance of this Agreement
pursuant to the Company’s instructions to the Optionee (including through an online acceptance process) is acceptable.
| Dated: |
|
|
|
| |
|
|
Optionee’s Signature |
| |
|
|
|
| |
|
|
Optionee’s name and address: |
| |
|
|
|
| |
|
|
|
| |
|
|
|
| |
|
|
|
| |
|
|
|
| |
|
|
|
NON-QUALIFIED
STOCK OPTION AGREEMENT
FOR COMPANY EMPLOYEES and consultants
UNDER the City Therapeutics, Inc.
2026 STOCK OPTION AND INCENTIVE PLAN
| Name of Optionee: |
____________________________________ |
| No. of Option Shares: |
___________________ |
| Option Exercise Price per Share: |
$__________________ |
[FMV on Grant
Date]
| Grant Date: |
__________________ |
| Expiration Date: |
__________________ |
Pursuant
to the City Therapeutics, Inc. 2026 Stock Option and Incentive Plan as amended through the date hereof (the “Plan”),
City Therapeutics, Inc. (the “Company”) hereby grants to the Optionee named above an option (the “Stock Option”)
to purchase on or prior to the Expiration Date specified above all or part of the number of shares of Common Stock, par value $0.0001
per share (the “Stock”) of the Company specified above at the Option Exercise Price per Share specified above subject to
the terms and conditions set forth herein and in the Plan. This Stock Option is not intended to be an “incentive stock option”
under Section 422 of the Internal Revenue Code of 1986, as amended.
1. Exercisability
Schedule. No portion of this Stock Option may be exercised until such portion shall have become exercisable. Except as set forth
below, and subject to the discretion of the Administrator (as defined in Section 2 of the Plan) to accelerate the exercisability
schedule hereunder, this Stock Option shall be exercisable as follows: ________________________, so long as the Optionee continues to
have a Service Relationship (as defined in the Plan) with the Company or a Subsidiary on such dates.
Once
exercisable, this Stock Option shall continue to be exercisable at any time or times prior to the close of business on the Expiration
Date, subject to the provisions hereof and of the Plan.
2. Manner
of Exercise.
(a) The
Optionee may exercise this Stock Option only in the following manner: from time to time on or prior to the Expiration Date of this Stock
Option, the Optionee may give written notice to the Administrator of his or her election to purchase some or all of the Option Shares
purchasable at the time of such notice. This notice shall specify the number of Option Shares to be purchased.
Payment
of the purchase price for the Option Shares may be made by one or more of the following methods: (i) in cash, by certified or bank
check or other instrument acceptable to the Administrator; (ii) through the delivery (or attestation to the ownership) of shares
of Stock that have been purchased by the Optionee on the open market or that are beneficially owned by the Optionee and are not then
subject to any restrictions under any Company plan and that otherwise satisfy any holding periods as may be required by the Administrator;
(iii) by the Optionee delivering to the Company a properly executed exercise notice together with irrevocable instructions to a
broker to promptly deliver to the Company cash or a check payable and acceptable to the Company to pay the option purchase price, provided
that in the event the Optionee chooses to pay the option purchase price as so provided, the Optionee and the broker shall comply with
such procedures and enter into such agreements of indemnity and other agreements as the Administrator shall prescribe as a condition
of such payment procedure; (iv) by a “net exercise” arrangement pursuant to which the Company will reduce the number
of shares of Stock issuable upon exercise by the largest whole number of shares with a Fair Market Value that does not exceed the aggregate
exercise price; or (v) a combination of (i), (ii), (iii) and (iv) above. Payment instruments will be received subject
to collection.
The
transfer to the Optionee on the records of the Company or of the transfer agent of the Option Shares will be contingent upon (i) the
Company’s receipt from the Optionee of the full purchase price for the Option Shares, as set forth above, (ii) the fulfillment
of any other requirements contained herein or in the Plan or in any other agreement or provision of laws, and (iii) the receipt
by the Company of any agreement, statement or other evidence that the Company may require to satisfy itself that the issuance of Stock
to be purchased pursuant to the exercise of Stock Options under the Plan and any subsequent resale of the shares of Stock will be in
compliance with applicable laws and regulations. In the event the Optionee chooses to pay the purchase price by previously-owned shares
of Stock through the attestation method, the number of shares of Stock transferred to the Optionee upon the exercise of the Stock Option
shall be net of the Shares attested to.
(b) The
shares of Stock purchased upon exercise of this Stock Option shall be transferred to the Optionee on the records of the Company or of
the transfer agent upon compliance to the satisfaction of the Administrator with all requirements under applicable laws or regulations
in connection with such transfer and with the requirements hereof and of the Plan. The determination of the Administrator as to such
compliance shall be final and binding on the Optionee. The Optionee shall not be deemed to be the holder of, or to have any of the rights
of a holder with respect to, any shares of Stock subject to this Stock Option unless and until this Stock Option shall have been exercised
pursuant to the terms hereof, the Company or the transfer agent shall have transferred the shares to the Optionee, and the Optionee’s
name shall have been entered as the stockholder of record on the books of the Company. Thereupon, the Optionee shall have full voting,
dividend and other ownership rights with respect to such shares of Stock.
(c) Notwithstanding
any other provision hereof or of the Plan, no portion of this Stock Option shall be exercisable after the Expiration Date hereof.
3. Termination
of Service Relationship. If the Optionee’s Service Relationship with the Company or a Subsidiary (as defined in the Plan) is
terminated, the period within which to exercise the Stock Option may be subject to earlier termination as set forth below.
(a) Termination
Due to Death. If the Optionee’s Service Relationship with the Company or a Subsidiary terminates by reason of the Optionee’s
death, any portion of this Stock Option outstanding on such date, to the extent exercisable on the date of death, may thereafter be exercised
by the Optionee’s legal representative or legatee for a period of twelve (12) months from the date of death or until the Expiration
Date, if earlier. Any portion of this Stock Option that is not exercisable on the date of death shall terminate immediately and be of
no further force or effect.
(b) Termination
Due to Disability. If the Optionee’s Service Relationship with the Company or a Subsidiary terminates by reason of the Optionee’s
disability (as determined by the Administrator), any portion of this Stock Option outstanding on such date, to the extent exercisable
on the date of such termination of Service Relationship, may thereafter be exercised by the Optionee for a period of twelve (12) months
from the date the Optionee’s Service Relationship is terminated by reason of the Optionee’s disability or until the Expiration
Date, if earlier. Any portion of this Stock Option that is not exercisable on the date of the termination of the Optionee’s Service
Relationship by reason of disability shall terminate immediately and be of no further force or effect.
(c) Termination
for Cause. If the Optionee’s Service Relationship with the Company or a Subsidiary terminates for Cause, any portion of this
Stock Option outstanding on such date shall terminate immediately and be of no further force and effect. For purposes hereof, “Cause”
shall mean, unless otherwise provided in an employment or other service agreement between the Company and the Optionee, a determination
by the Administrator that the Optionee shall be dismissed as a result of (i) any material breach by the Optionee of any agreement
between the Optionee and the Company; (ii) the conviction of, indictment for or plea of nolo contendere by the Optionee to a felony
or a crime involving moral turpitude; or (iii) any material misconduct or willful and deliberate non-performance (other than by
reason of disability) by the Optionee of the Optionee’s duties to the Company.
(d) Other
Termination. If the Optionee’s Service Relationship with the Company or a Subsidiary terminates for any reason other than the
Optionee’s death, the Optionee’s disability or Cause, and unless otherwise determined by the Administrator, any portion of
this Stock Option outstanding on such date may be exercised, to the extent exercisable on the date of termination, for a period of ninety
(90) days from the date of termination or until the Expiration Date, if earlier. Any portion of this Stock Option that is not exercisable
on the date of termination shall terminate immediately and be of no further force or effect.
The
Administrator’s determination of the reason for termination of the Optionee’s Service Relationship with the Company or a
Subsidiary shall be conclusive and binding on the Optionee and his or her representatives or legatees.
4. Incorporation
of Plan. Notwithstanding anything herein to the contrary, this Stock Option shall be subject to and governed by all the terms and
conditions of the Plan, including the powers of the Administrator set forth in Section 2(b) of the Plan. Capitalized terms
in this Agreement shall have the meaning specified in the Plan, unless a different meaning is specified herein.
5. Transferability.
This Agreement is personal to the Optionee, is non-assignable and is not transferable in any manner, by operation of law or otherwise,
other than by will or the laws of descent and distribution. This Stock Option is exercisable, during the Optionee’s lifetime, only
by the Optionee, and thereafter, only by the Optionee’s legal representative or legatee.
6. Tax
Withholding. The Optionee shall, not later than the date as of which the exercise of this Stock Option becomes a taxable event for
Federal income tax purposes, pay to the Company or make arrangements satisfactory to the Administrator for payment of any Federal, state
and local taxes required by law to be withheld on account of such taxable event. The Company shall have the authority to cause the required
tax withholding obligation to be satisfied, in whole or in part, by (i) withholding from shares of Stock to be issued to the Optionee
a number of shares of Stock with an aggregate Fair Market Value that would satisfy the withholding amount due; or (ii) causing its
transfer agent to sell from the number of shares of Stock to be issued to the Optionee, the number of shares of Stock necessary to satisfy
the Federal, state and local taxes required by law to be withheld from the Optionee on account of such transfer.
7. No
Obligation to Continue Service Relationship. Neither the Company nor any Subsidiary is obligated by or as a result of the Plan or
this Agreement to continue the Optionee in a Service Relationship with the Company or a Subsidiary and neither the Plan nor this Agreement
shall interfere in any way with the right of the Company or any Subsidiary to terminate the Optionee’s Service Relationship with
the Company or a Subsidiary at any time.
8. Integration.
This Agreement constitutes the entire agreement between the parties with respect to this Stock Option and supersedes all prior agreements
and discussions between the parties concerning such subject matter.
9. Data
Privacy Consent. In order to administer the Plan and this Agreement and to implement or structure future equity grants, the Company,
its subsidiaries and affiliates and certain agents thereof (together, the “Relevant Companies”) may process any and all personal
or professional data, including but not limited to, Social Security or other identification number, home address and telephone number,
date of birth and other information that is necessary or desirable for the administration of the Plan and/or this Agreement (the “Relevant
Information”). By entering into this Agreement, the Optionee (i) authorizes the Company to collect, process, register and
transfer to the Relevant Companies all Relevant Information; (ii) waives any privacy rights the Optionee may have with respect to
the Relevant Information; (iii) authorizes the Relevant Companies to store and transmit such information in electronic form; and
(iv) authorizes the transfer of the Relevant Information to any jurisdiction in which the Relevant Companies consider appropriate.
The Optionee shall have access to, and the right to change, the Relevant Information. Relevant Information will only be used in accordance
with applicable law.
10. Notices.
Notices hereunder shall be mailed or delivered to the Company at its principal place of business and shall be mailed or delivered to
the Optionee at the address on file with the Company or, in either case, at such other address as one party may subsequently furnish
to the other party in writing.
11. [Clawback
Acknowledgement. The Optionee acknowledges that the Optionee may become subject to the City Therapeutics, Inc. Compensation
Recovery Policy adopted pursuant to Rule 10D-1 promulgated under the Exchange Act and NASDAQ Rule 5608, or any successor rule (the
“Clawback Policy”). The Optionee understands that if the Optionee is or becomes subject to the Clawback Policy, the Company
and/or the Board shall be entitled to recover all Erroneously Awarded Compensation (as defined in the Clawback Policy) from the Optionee
pursuant to such means as the Company and/or the Board may elect. The Optionee agrees that the Optionee shall take all required action
to enable such recovery. The Optionee understands that such recovery may be sought and occur after the Optionee’s employment or
service with the Company terminates. The Optionee further agrees that the Optionee is not entitled to indemnification for any Erroneously
Awarded Compensation or for any claim or losses arising out of or in any way related to Erroneously Awarded Compensation recovered pursuant
to the Clawback Policy and, to the extent any agreement or organizational document purports to provide otherwise, the Optionee hereby
irrevocably agrees to forego such indemnification. The Optionee acknowledges and agrees that the Optionee has received and has had an
opportunity to review the Clawback Policy. Any action by the Company to recover Erroneously Awarded Compensation under the Clawback Policy
from the Optionee shall not, whether alone or in combination with any other action, event or condition, be deemed (i) an event giving
rise to a right to resign for Good Reason (as defined in the City Therapeutics, Inc. Executive Severance Plan) or serve as a basis
for a claim of constructive termination under any benefits or compensation arrangement applicable to the Optionee, or (ii) to constitute
a breach of a contract or other arrangement to which the Optionee is a party. This Section 11 is a material term of this Agreement.]1
1
Note to Draft: For Section 16 officers only.
| |
City Therapeutics, Inc. |
| |
|
|
| |
By: |
|
| |
|
Title: |
The foregoing Agreement
is hereby accepted and the terms and conditions thereof hereby agreed to by the undersigned. Electronic acceptance of this Agreement
pursuant to the Company’s instructions to the Optionee (including through an online acceptance process) is acceptable.
| Dated: |
|
|
|
| |
|
|
Optionee’s Signature |
| |
|
|
|
| |
|
|
Optionee’s name and address: |
| |
|
|
|
| |
|
|
|
| |
|
|
|
| |
|
|
|
| |
|
|
|
| |
|
|
|
NON-QUALIFIED
STOCK OPTION AGREEMENT
FOR Company NON-EMPLOYEE DIRECTORS
UNDER The City Therapeutics, Inc.
2026 STOCK OPTION AND INCENTIVE PLAN
| Name of Optionee: |
____________________________________ |
| No. of Option Shares: |
___________________ |
| Option Exercise Price per Share: |
$__________________ |
[FMV on Grant
Date]
| Grant Date: |
__________________ |
| Expiration Date: |
__________________ |
[No more than 10 years]
Pursuant
to the City Therapeutics, Inc. 2026 Stock Option and Incentive Plan as amended through the date hereof (the “Plan”),
City Therapeutics, Inc. (the “Company”) hereby grants to the Optionee named above, who is a Non-Employee Director of
the Company but is not an employee of the Company, an option (the “Stock Option”) to purchase on or prior to the Expiration
Date specified above all or part of the number of shares of Common Stock, par value $0.0001 per share (the “Stock”), of the
Company specified above at the Option Exercise Price per Share specified above subject to the terms and conditions set forth herein and
in the Plan. This Stock Option is not intended to be an “incentive stock option” under Section 422 of the Internal Revenue
Code of 1986, as amended.
1. Exercisability
Schedule. No portion of this Stock Option may be exercised until such portion shall have become exercisable. Except as set forth
below, and subject to the discretion of the Administrator (as defined in Section 2 of the Plan) to accelerate the exercisability
schedule hereunder, this Stock Option shall be exercisable as follows: ________________________, so long as the Optionee remains in service
as a member of the Board on such dates.
Notwithstanding
the foregoing, in the event of a Sale Event, 100% of the then-outstanding and unvested Option Shares shall immediately be deemed vested
and exercisable on the date of such Sale Event; provided that the Optionee remains in service as a member of the Board until the date
of such Sale Event. Once exercisable, this Stock Option shall continue to be exercisable at any time or times prior to the close of business
on the Expiration Date, subject to the provisions hereof and of the Plan.
2. Manner
of Exercise.
(a) The
Optionee may exercise this Stock Option only in the following manner: from time to time on or prior to the Expiration Date of this Stock
Option, the Optionee may give written notice to the Administrator of his or her election to purchase some or all of the Option Shares
purchasable at the time of such notice. This notice shall specify the number of Option Shares to be purchased.
Payment
of the purchase price for the Option Shares may be made by one or more of the following methods: (i) in cash, by certified or bank
check or other instrument acceptable to the Administrator; (ii) through the delivery (or attestation to the ownership) of shares
of Stock that have been purchased by the Optionee on the open market or that are beneficially owned by the Optionee and are not then
subject to any restrictions under any Company plan and that otherwise satisfy any holding periods as may be required by the Administrator;
(iii) by the Optionee delivering to the Company a properly executed exercise notice together with irrevocable instructions to a
broker to promptly deliver to the Company cash or a check payable and acceptable to the Company to pay the option purchase price, provided
that in the event the Optionee chooses to pay the option purchase price as so provided, the Optionee and the broker shall comply with
such procedures and enter into such agreements of indemnity and other agreements as the Administrator shall prescribe as a condition
of such payment procedure; (iv) by a “net exercise” arrangement pursuant to which the Company will reduce the number
of shares of Stock issuable upon exercise by the largest whole number of shares with a Fair Market Value that does not exceed the aggregate
exercise price; or (v) a combination of (i), (ii), (iii) and (iv) above. Payment instruments will be received subject
to collection.
The
transfer to the Optionee on the records of the Company or of the transfer agent of the Option Shares will be contingent upon (i) the
Company’s receipt from the Optionee of the full purchase price for the Option Shares, as set forth above, (ii) the fulfillment
of any other requirements contained herein or in the Plan or in any other agreement or provision of laws and (iii) the receipt by
the Company of any agreement, statement or other evidence that the Company may require to satisfy itself that the issuance of Stock to
be purchased pursuant to the exercise of Stock Options under the Plan and any subsequent resale of the shares of Stock will be in compliance
with applicable laws and regulations. In the event the Optionee chooses to pay the purchase price by previously-owned shares of Stock
through the attestation method, the number of shares of Stock transferred to the Optionee upon the exercise of the Stock Option shall
be net of the Shares attested to.
(b) The
shares of Stock purchased upon exercise of this Stock Option shall be transferred to the Optionee on the records of the Company or of
the transfer agent upon compliance to the satisfaction of the Administrator with all requirements under applicable laws or regulations
in connection with such transfer and with the requirements hereof and of the Plan. The determination of the Administrator as to such
compliance shall be final and binding on the Optionee. The Optionee shall not be deemed to be the holder of, or to have any of the rights
of a holder with respect to, any shares of Stock subject to this Stock Option unless and until this Stock Option shall have been exercised
pursuant to the terms hereof, the Company or the transfer agent shall have transferred the shares to the Optionee, and the Optionee’s
name shall have been entered as the stockholder of record on the books of the Company. Thereupon, the Optionee shall have full voting,
dividend and other ownership rights with respect to such shares of Stock.
(c) Notwithstanding
any other provision hereof or of the Plan, no portion of this Stock Option shall be exercisable after the Expiration Date hereof.
3. Termination
as Non-Employee Director. If the Optionee ceases to be a Non-Employee Director of the Company, the period within which to exercise
the Stock Option may be subject to earlier termination as set forth below.
(a) Termination
Due to Death. If the Optionee’s service as a Non-Employee Director terminates by reason of the Optionee’s death, any
portion of this Stock Option outstanding on such date, to the extent exercisable on the date of death, may thereafter be exercised by
the Optionee’s legal representative or legatee for a period of twelve (12) months from the date of death or until the Expiration
Date, if earlier. Any portion of this Stock Option that is not exercisable on the date of death shall terminate immediately and be of
no further force or effect.
(b) Termination
Due to Disability. If the Optionee’s Service Relationship (as defined in the Plan) with the Company or a Subsidiary terminates
by reason of the Optionee’s disability (as determined by the Administrator), any portion of this Stock Option outstanding on such
date, to the extent exercisable on the date of such termination, may thereafter be exercised by the Optionee for a period of twelve (12)
months from the date the Optionee’s Service Relationship is terminated by reason of the Optionee’s disability or until the
Expiration Date, if earlier. Any portion of this Stock Option that is not exercisable on the date of the termination of the Optionee’s
Service Relationship by reason of disability shall terminate immediately and be of no further force or effect.
(c) Termination
for Cause. If the Optionee’s Service Relationship with the Company or a Subsidiary terminates for Cause, any portion of this
Stock Option outstanding on such date shall terminate immediately and be of no further force and effect. For purposes hereof, “Cause”
shall mean, unless otherwise provided in an employment or other service agreement between the Company and the Optionee, a determination
by the Administrator that the Optionee shall be dismissed as a result of (i) any material breach by the Optionee of any agreement
between the Optionee and the Company; (ii) the conviction of, indictment for or plea of nolo contendere by the Optionee to a felony
or a crime involving moral turpitude; or (iii) any material misconduct or willful and deliberate non-performance (other than by
reason of disability) by the Optionee of the Optionee’s duties to the Company.
(d) Other
Termination. If the Optionee ceases to be a Non-Employee Director for any reason other than the Optionee’s death, the Optionee’s
disability or Cause, and unless otherwise determined by the Administrator, any portion of this Stock Option outstanding on such date
may be exercised, to the extent exercisable on the date the Optionee ceased to be a Non-Employee Director, for a period of twelve (12)
months from the date the Optionee ceased to be a Non-Employee Director or until the Expiration Date, if earlier. Any portion of this
Stock Option that is not exercisable on the date the Optionee ceases to be a Non-Employee Director shall terminate immediately and be
of no further force or effect.
4. Incorporation
of Plan. Notwithstanding anything herein to the contrary, this Stock Option shall be subject to and governed by all the terms and
conditions of the Plan, including the powers of the Administrator set forth in Section 2(b) of the Plan. Capitalized terms
in this Agreement shall have the meaning specified in the Plan, unless a different meaning is specified herein.
5. Transferability.
This Agreement is personal to the Optionee, is non-assignable and is not transferable in any manner, by operation of law or otherwise,
other than by will or the laws of descent and distribution. This Stock Option is exercisable, during the Optionee’s lifetime, only
by the Optionee, and thereafter, only by the Optionee’s legal representative or legatee.
6. No
Obligation to Continue Service Relationship. Neither the Company nor any Subsidiary is obligated by or as a result of the Plan or
this Agreement to continue the Optionee in a Service Relationship with the Company or a Subsidiary and neither the Plan nor this Agreement
shall interfere in any way with the right of the Company or any Subsidiary to terminate the Optionee’s Service Relationship with
the Company or a Subsidiary at any time.
7. Integration.
This Agreement constitutes the entire agreement between the parties with respect to this Stock Option and supersedes all prior agreements
and discussions between the parties concerning such subject matter.
8. Data
Privacy Consent. In order to administer the Plan and this Agreement and to implement or structure future equity grants, the Company,
its subsidiaries and affiliates and certain agents thereof (together, the “Relevant Companies”) may process any and all personal
or professional data, including but not limited to Social Security or other identification number, home address and telephone number,
date of birth and other information that is necessary or desirable for the administration of the Plan and/or this Agreement (the “Relevant
Information”). By entering into this Agreement, the Optionee (i) authorizes the Company to collect, process, register and
transfer to the Relevant Companies all Relevant Information; (ii) waives any privacy rights the Optionee may have with respect to
the Relevant Information; (iii) authorizes the Relevant Companies to store and transmit such information in electronic form; and
(iv) authorizes the transfer of the Relevant Information to any jurisdiction in which the Relevant Companies consider appropriate.
The Optionee shall have access to, and the right to change, the Relevant Information. Relevant Information will only be used in accordance
with applicable law.
9. Notices.
Notices hereunder shall be mailed or delivered to the Company at its principal place of business and shall be mailed or delivered to
the Optionee at the address on file with the Company or, in either case, at such other address as one party may subsequently furnish
to the other party in writing.
| |
City Therapeutics, Inc. |
| |
|
|
| |
By: |
|
| |
|
Title: |
The foregoing Agreement
is hereby accepted and the terms and conditions thereof hereby agreed to by the undersigned. Electronic acceptance of this Agreement
pursuant to the Company’s instructions to the Optionee (including through an online acceptance process) is acceptable.
| Dated: |
|
|
|
| |
|
|
Optionee’s Signature |
| |
|
|
|
| |
|
|
Optionee’s name and address: |
| |
|
|
|
| |
|
|
|
| |
|
|
|
| |
|
|
|
| |
|
|
|
| |
|
|
|
RESTRICTED
STOCK AWARD AGREEMENT
UNDER THE City Therapeutics, Inc.
2026 STOCK OPTION AND INCENTIVE PLAN
| Name of Grantee: |
____________________________________ |
| No. of Shares: |
__________________ |
| Grant Date: |
__________________ |
Pursuant
to the City Therapeutics, Inc. 2026 Stock Option and Incentive Plan (the “Plan”) as amended through the date hereof,
City Therapeutics, Inc. (the “Company”) hereby grants a Restricted Stock Award (an “Award”) to the Grantee
named above. Upon acceptance of this Award, the Grantee shall receive the number of shares of Common Stock, par value $0.0001 per share
(the “Stock”) of the Company specified above, subject to the restrictions and conditions set forth herein and in the Plan.
The Company acknowledges the receipt from the Grantee of consideration with respect to the par value of the Stock in the form of cash,
past or future services rendered to the Company by the Grantee or such other form of consideration as is acceptable to the Administrator.
1. Award.
The shares of Restricted Stock awarded hereunder shall be issued and held by the Company’s transfer agent in book entry form, and
the Grantee’s name shall be entered as the stockholder of record on the books of the Company. Thereupon, the Grantee shall have
all the rights of a stockholder with respect to such shares, including voting and dividend rights, subject, however, to the restrictions
and conditions specified in Paragraph 2 below. The Grantee shall (i) sign and deliver to the Company a copy of this Award Agreement
and (ii) deliver to the Company a stock power endorsed in blank.
2. Restrictions
and Conditions.
(a) Any
book entries for the shares of Restricted Stock granted herein shall bear an appropriate legend, as determined by the Administrator in
its sole discretion, to the effect that such shares are subject to restrictions as set forth herein and in the Plan.
(b) Shares
of Restricted Stock granted herein may not be sold, assigned, transferred, pledged or otherwise encumbered or disposed of by the Grantee
prior to vesting.
(c) If
the Grantee’s Service Relationship (as defined in the Plan) with the Company or a Subsidiary is voluntarily or involuntarily terminated
for any reason (including due to death or disability) prior to vesting of shares of Restricted Stock granted herein, all shares of Restricted
Stock shall immediately and automatically be forfeited and returned to the Company.
3. Vesting
of Restricted Stock. The restrictions and conditions in Paragraph 2 of this Agreement shall lapse as follows: _________________________
(each such date, a “Vesting Date”), so long as the Grantee continues to have a Service Relationship with the Company or a
Subsidiary on such Vesting Dates. If a series of Vesting Dates is specified, then the restrictions and conditions in Paragraph 2
shall lapse only with respect to the number of shares of Restricted Stock specified as vested on such date.
Subsequent
to such Vesting Date or Dates, the shares of Stock on which all restrictions and conditions have lapsed shall no longer be deemed Restricted
Stock. The Administrator may at any time accelerate the vesting schedule specified in this Paragraph 3.
4. Dividends.
Dividends on shares of Restricted Stock shall be paid currently to the Grantee.
5. Incorporation
of Plan. Notwithstanding anything herein to the contrary, this Award shall be subject to and governed by all the terms and conditions
of the Plan, including the powers of the Administrator set forth in Section 2(b) of the Plan. Capitalized terms in this Agreement
shall have the meaning specified in the Plan, unless a different meaning is specified herein.
6. Transferability.
This Agreement is personal to the Grantee, is non-assignable and is not transferable in any manner, by operation of law or otherwise,
other than by will or the laws of descent and distribution.
7. Tax
Withholding. The Grantee shall, not later than the date as of which the receipt of this Award becomes a taxable event for Federal
income tax purposes, pay to the Company or make arrangements satisfactory to the Administrator for payment of any Federal, state and
local taxes required by law to be withheld on account of such taxable event. Except in the case where an election is made pursuant to
Paragraph 8 below, the Company shall have the authority to cause the required tax withholding obligation to be satisfied, in whole or
in part, by (i) withholding from shares of Stock to be issued or released by the transfer agent a number of shares of Stock with
an aggregate Fair Market Value that would satisfy the withholding amount due; or (ii) causing its transfer agent to sell from the
number of shares of Stock to be issued or released to the Grantee, the number of shares of Stock necessary to satisfy the Federal, state
and local taxes required by law to be withheld from the Grantee on account of such transfer.
8. Election
Under Section 83(b). The Grantee and the Company hereby agree that the Grantee may, within 30 days following the Grant Date
of this Award, file with the Internal Revenue Service and the Company an election under Section 83(b) of the Internal Revenue
Code. In the event the Grantee makes such an election, he or she agrees to provide a copy of the election to the Company. The Grantee
acknowledges that he or she is responsible for obtaining the advice of his or her tax advisors with regard to the Section 83(b) election
and that he or she is relying solely on such advisors and not on any statements or representations of the Company or any of its agents
with regard to such election.
9. No
Obligation to Continue Service Relationship. Neither the Company nor any Subsidiary is obligated by or as a result of the Plan or
this Agreement to continue the Grantee in a Service Relationship with the Company or a Subsidiary and neither the Plan nor this Agreement
shall interfere in any way with the right of the Company or any Subsidiary to terminate the Grantee’s Service Relationship with
the Company or a Subsidiary at any time.
10. Integration.
This Agreement constitutes the entire agreement between the parties with respect to this Award and supersedes all prior agreements and
discussions between the parties concerning such subject matter.
11. Data
Privacy Consent. In order to administer the Plan and this Agreement and to implement or structure future equity grants, the Company,
its subsidiaries and affiliates and certain agents thereof (together, the “Relevant Companies”) may process any and all personal
or professional data, including but not limited to, Social Security or other identification number, home address and telephone number,
date of birth and other information that is necessary or desirable for the administration of the Plan and/or this Agreement (the “Relevant
Information”). By entering into this Agreement, the Grantee (i) authorizes the Company to collect, process, register and transfer
to the Relevant Companies all Relevant Information; (ii) waives any privacy rights the Grantee may have with respect to the Relevant
Information; (iii) authorizes the Relevant Companies to store and transmit such information in electronic form; and (iv) authorizes
the transfer of the Relevant Information to any jurisdiction in which the Relevant Companies consider appropriate. The Grantee shall
have access to, and the right to change, the Relevant Information. Relevant Information will only be used in accordance with applicable
law.
12. Notices.
Notices hereunder shall be mailed or delivered to the Company at its principal place of business and shall be mailed or delivered to
the Grantee at the address on file with the Company or, in either case, at such other address as one party may subsequently furnish to
the other party in writing.
13. [Clawback
Acknowledgement. The Grantee acknowledges that the Grantee may become subject to the City Therapeutics, Inc. Compensation Recovery
Policy adopted pursuant to Rule 10D-1 promulgated under the Exchange Act and NASDAQ Rule 5608, or any successor rule (the
“Clawback Policy”). The Grantee understands that if the Grantee is or becomes subject to the Clawback Policy, the Company
and/or the Board shall be entitled to recover all Erroneously Awarded Compensation (as defined in the Clawback Policy) from the Grantee
pursuant to such means as the Company and/or the Board may elect. The Grantee agrees that the Grantee shall take all required action
to enable such recovery. The Grantee understands that such recovery may be sought and occur after the Grantee’s employment or service
with the Company terminates. The Grantee further agrees that the Grantee is not entitled to indemnification for any Erroneously Awarded
Compensation or for any claim or losses arising out of or in any way related to Erroneously Awarded Compensation recovered pursuant to
the Clawback Policy and, to the extent any agreement or organizational document purports to provide otherwise, the Grantee hereby irrevocably
agrees to forego such indemnification. The Grantee acknowledges and agrees that the Grantee has received and has had an opportunity to
review the Clawback Policy. Any action by the Company to recover Erroneously Awarded Compensation under the Clawback Policy from the
Grantee shall not, whether alone or in combination with any other action, event or condition, be deemed (i) an event giving rise
to a right to resign for Good Reason (as defined in the City Therapeutics, Inc. Executive Severance Plan) or serve as a basis for
a claim of constructive termination under any benefits or compensation arrangement applicable to the Grantee, or (ii) to constitute
a breach of a contract or other arrangement to which the Grantee is a party. This Section 13 is a material term of this Agreement.]1
1
Note to Draft: For Section 16 officers only.
| |
City Therapeutics, Inc. |
| |
|
|
| |
By: |
|
| |
|
Title: |
The foregoing Agreement
is hereby accepted and the terms and conditions thereof hereby agreed to by the undersigned. Electronic acceptance of this Agreement
pursuant to the Company’s instructions to the Grantee (including through an online acceptance process) is acceptable.
| Dated: |
|
|
|
| |
|
|
Grantee’s Signature |
| |
|
|
|
| |
|
|
Grantee’s name and address: |
| |
|
|
|
| |
|
|
|
| |
|
|
|
| |
|
|
|
| |
|
|
|
| |
|
|
|
RESTRICTED
STOCK UNIT AWARD AGREEMENT
FOR COMPANY EMPLOYEES and consultants
UNDER the City Therapeutics, Inc.
2026 STOCK OPTION AND INCENTIVE PLAN
| Name of Grantee: |
____________________________________ |
| No. of Restricted Stock Units: |
__________________ |
| Grant Date: |
__________________ |
Pursuant
to the City Therapeutics, Inc. 2026 Stock Option and Incentive Plan as amended through the date hereof (the “Plan”),
City Therapeutics, Inc. (the “Company”) hereby grants an award of the number of Restricted Stock Units listed above
(an “Award”) to the Grantee named above. Each Restricted Stock Unit shall relate to one share of Common Stock, par value
$0.0001 per share (the “Stock”) of the Company.
1. Restrictions
on Transfer of Award. This Award may not be sold, transferred, pledged, assigned or otherwise encumbered or disposed of by the Grantee,
and any shares of Stock issuable with respect to the Award may not be sold, transferred, pledged, assigned or otherwise encumbered or
disposed of until (i) the Restricted Stock Units have vested as provided in Paragraph 2 of this Agreement and (ii) shares of
Stock have been issued to the Grantee in accordance with the terms of the Plan and this Agreement.
2. Vesting
of Restricted Stock Units. The restrictions and conditions of Paragraph 1 of this Agreement shall lapse as follows: ______________________
(each such date, a “Vesting Date”), so long as the Grantee continues to have a Service Relationship (as defined in the Plan)
with the Company or a Subsidiary on such Vesting Dates. If a series of Vesting Dates is specified, then the restrictions and conditions
in Paragraph 1 shall lapse only with respect to the number of Restricted Stock Units specified as vested on such date.
The
Administrator may at any time accelerate the vesting schedule specified in this Paragraph 2.
3. Termination
of Service Relationship. If the Grantee’s Service Relationship with the Company or a Subsidiary terminates for any reason (including
death or disability) prior to the satisfaction of the vesting conditions set forth in Paragraph 2 above, any Restricted Stock Units that
have not vested as of such date shall automatically and without notice terminate and be forfeited, and neither the Grantee nor any of
his or her successors, heirs, assigns or personal representatives will thereafter have any further rights or interests in such unvested
Restricted Stock Units.
4. Issuance
of Shares of Stock. As soon as practicable following each Vesting Date (but in no event later than two and one-half months after
the end of the year in which the Vesting Date occurs), the Company shall issue to the Grantee the number of shares of Stock equal to
the aggregate number of Restricted Stock Units that have vested pursuant to Paragraph 2 of this Agreement on such date and the Grantee
shall thereafter have all the rights of a stockholder of the Company with respect to such shares.
5. Incorporation
of Plan. Notwithstanding anything herein to the contrary, this Agreement shall be subject to and governed by all the terms and conditions
of the Plan, including the powers of the Administrator set forth in Section 2(b) of the Plan. Capitalized terms in this Agreement
shall have the meaning specified in the Plan, unless a different meaning is specified herein.
6. Tax
Withholding. The Grantee shall, not later than the date as of which the receipt of this Award becomes a taxable event for Federal
income tax purposes, pay to the Company or make arrangements satisfactory to the Administrator for payment of any Federal, state and
local taxes required by law to be withheld on account of such taxable event. The Company shall have the authority to cause the required
tax withholding obligation to be satisfied, in whole or in part, by (i) withholding from shares of Stock to be issued to the Grantee
a number of shares of Stock with an aggregate Fair Market Value that would satisfy the withholding amount due; or (ii) causing its
transfer agent to sell from the number of shares of Stock to be issued to the Grantee, the number of shares of Stock necessary to satisfy
the Federal, state and local taxes required by law to be withheld from the Grantee on account of such transfer.
7. Section 409A
of the Code. This Agreement shall be interpreted in such a manner that all provisions relating to the settlement of the Award are
exempt from the requirements of Section 409A of the Code as “short-term deferrals” as described in Section 409A
of the Code.
8. No
Obligation to Continue Service Relationship. Neither the Company nor any Subsidiary is obligated by or as a result of the Plan or
this Agreement to continue the Grantee in a Service Relationship with the Company or a Subsidiary and neither the Plan nor this Agreement
shall interfere in any way with the right of the Company or any Subsidiary to terminate the Grantee’s Service Relationship with
the Company or a Subsidiary at any time.
9. Integration.
This Agreement constitutes the entire agreement between the parties with respect to this Award and supersedes all prior agreements and
discussions between the parties concerning such subject matter.
10. Data
Privacy Consent. In order to administer the Plan and this Agreement and to implement or structure future equity grants, the Company,
its subsidiaries and affiliates and certain agents thereof (together, the “Relevant Companies”) may process any and all personal
or professional data, including but not limited to, Social Security or other identification number, home address and telephone number,
date of birth and other information that is necessary or desirable for the administration of the Plan and/or this Agreement (the “Relevant
Information”). By entering into this Agreement, the Grantee (i) authorizes the Company to collect, process, register and transfer
to the Relevant Companies all Relevant Information; (ii) waives any privacy rights the Grantee may have with respect to the Relevant
Information; (iii) authorizes the Relevant Companies to store and transmit such information in electronic form; and (iv) authorizes
the transfer of the Relevant Information to any jurisdiction in which the Relevant Companies consider appropriate. The Grantee shall
have access to, and the right to change, the Relevant Information. Relevant Information will only be used in accordance with applicable
law.
11. Notices.
Notices hereunder shall be mailed or delivered to the Company at its principal place of business and shall be mailed or delivered to
the Grantee at the address on file with the Company or, in either case, at such other address as one party may subsequently furnish to
the other party in writing.
12. [Clawback
Acknowledgement. The Grantee acknowledges that the Grantee may become subject to the City Therapeutics, Inc. Compensation Recovery
Policy adopted pursuant to Rule 10D-1 promulgated under the Exchange Act and Nasdaq Rule 5608, or any successor rule (the
“Clawback Policy”). The Grantee understands that if the Grantee is or becomes subject to the Clawback Policy, the Company
and/or the Board shall be entitled to recover all Erroneously Awarded Compensation (as defined in the Clawback Policy) from the Grantee
pursuant to such means as the Company and/or the Board may elect. The Grantee agrees that the Grantee shall take all required action
to enable such recovery. The Grantee understands that such recovery may be sought and occur after the Grantee’s employment or service
with the Company terminates. The Grantee further agrees that the Grantee is not entitled to indemnification for any Erroneously Awarded
Compensation or for any claim or losses arising out of or in any way related to Erroneously Awarded Compensation recovered pursuant to
the Clawback Policy and, to the extent any agreement or organizational document purports to provide otherwise, the Grantee hereby irrevocably
agrees to forego such indemnification. The Grantee acknowledges and agrees that the Grantee has received and has had an opportunity to
review the Clawback Policy. Any action by the Company to recover Erroneously Awarded Compensation under the Clawback Policy from the
Grantee shall not, whether alone or in combination with any other action, event or condition, be deemed (i) an event giving rise
to a right to resign for Good Reason (as defined in the City Therapeutics, Inc. Executive Severance Plan) or serve as a basis for
a claim of constructive termination under any benefits or compensation arrangement applicable to the Grantee, or (ii) to constitute
a breach of a contract or other arrangement to which the Grantee is a party. This Section 12 is a material term of this Agreement.]1
| |
City Therapeutics, Inc. |
| |
|
|
| |
By: |
|
| |
|
Title: |
The foregoing Agreement
is hereby accepted and the terms and conditions thereof hereby agreed to by the undersigned. Electronic acceptance of this Agreement
pursuant to the Company’s instructions to the Grantee (including through an online acceptance process) is acceptable.
| Dated: |
|
|
|
| |
|
|
Grantee’s Signature |
| |
|
|
|
| |
|
|
Grantee’s name and
address: |
| |
|
|
|
| |
|
|
|
| |
|
|
|
| |
|
|
|
| |
|
|
|
| |
|
|
|
1
Note to Draft: For Section 16 officers only.
RESTRICTED
STOCK UNIT AWARD AGREEMENT
FOR NON-EMPLOYEE DIRECTORS
UNDER the City Therapeutics, Inc.
2026 STOCK OPTION AND INCENTIVE PLAN
| Name of Grantee: |
____________________________________ |
| No. of Restricted Stock Units: |
__________________ |
| Grant Date: |
__________________ |
Pursuant
to the City Therapeutics, Inc. 2026 Stock Option and Incentive Plan as amended through the date hereof (the “Plan”),
City Therapeutics, Inc. (the “Company”) hereby grants an award of the number of Restricted Stock Units listed above
(an “Award”) to the Grantee named above. Each Restricted Stock Unit shall relate to one share of Common Stock, par value
$0.0001 per share (the “Stock”) of the Company.
1. Restrictions
on Transfer of Award. This Award may not be sold, transferred, pledged, assigned or otherwise encumbered or disposed of by the Grantee,
and any shares of Stock issuable with respect to the Award may not be sold, transferred, pledged, assigned or otherwise encumbered or
disposed of until (i) the Restricted Stock Units have vested as provided in Paragraph 2 of this Agreement and (ii) shares of
Stock have been issued to the Grantee in accordance with the terms of the Plan and this Agreement.
2. Vesting
of Restricted Stock Units. The restrictions and conditions of Paragraph 1 of this Agreement shall lapse as follows: ___________________
(each such date, a “Vesting Date”), so long as the Grantee remains in service as a member of the Board on such Vesting Dates.
If a series of Vesting Dates is specified, then the restrictions and conditions in Paragraph 1 shall lapse only with respect to
the number of Restricted Stock Units specified as vested on such date.
Notwithstanding
the foregoing, in the event of a Sale Event, 100% of the then-outstanding and unvested Restricted Stock Units shall immediately be deemed
vested on the date of such Sale Event; provided that the Grantee remains in service as a member of the Board until the date of such Sale
Event. The Administrator may at any time accelerate the vesting schedule specified in this Paragraph 2.
3. Termination
of Service as a Non-Employee Director. If the Grantee’s service as a Non-Employee Director terminates for any reason (including
death or disability) prior to the satisfaction of the vesting conditions set forth in Paragraph 2 above, any Restricted Stock Units that
have not vested as of such date shall automatically and without notice terminate and be forfeited, and neither the Grantee nor any of
his or her successors, heirs, assigns or personal representatives will thereafter have any further rights or interests in such unvested
Restricted Stock Units.
4. Issuance
of Shares of Stock. As soon as practicable following each Vesting Date (but in no event later than two and one-half months after
the end of the year in which the Vesting Date occurs), the Company shall issue to the Grantee the number of shares of Stock equal to
the aggregate number of Restricted Stock Units that have vested pursuant to Paragraph 2 of this Agreement on such date and the Grantee
shall thereafter have all the rights of a stockholder of the Company with respect to such shares.
5. Incorporation
of Plan. Notwithstanding anything herein to the contrary, this Agreement shall be subject to and governed by all the terms and conditions
of the Plan, including the powers of the Administrator set forth in Section 2(b) of the Plan. Capitalized terms in this Agreement
shall have the meaning specified in the Plan, unless a different meaning is specified herein.
6. Section 409A
of the Code. This Agreement shall be interpreted in such a manner that all provisions relating to the settlement of the Award are
exempt from the requirements of Section 409A of the Code as “short-term deferrals” as described in Section 409A
of the Code.
7. No
Obligation to Continue Service Relationship. Neither the Company nor any Subsidiary is obligated by or as a result of the Plan or
this Agreement to continue the Grantee in a Service Relationship (as defined in the Plan) with the Company or a Subsidiary and neither
the Plan nor this Agreement shall interfere in any way with the right of the Company or any Subsidiary to terminate the Grantee’s
Service Relationship with the Company or a Subsidiary at any time.
8. Integration.
This Agreement constitutes the entire agreement between the parties with respect to this Award and supersedes all prior agreements and
discussions between the parties concerning such subject matter.
9. Data
Privacy Consent. In order to administer the Plan and this Agreement and to implement or structure future equity grants, the Company,
its subsidiaries and affiliates and certain agents thereof (together, the “Relevant Companies”) may process any and all personal
or professional data, including but not limited to, Social Security or other identification number, home address and telephone number,
date of birth and other information that is necessary or desirable for the administration of the Plan and/or this Agreement (the “Relevant
Information”). By entering into this Agreement, the Grantee (i) authorizes the Company to collect, process, register and transfer
to the Relevant Companies all Relevant Information; (ii) waives any privacy rights the Grantee may have with respect to the Relevant
Information; (iii) authorizes the Relevant Companies to store and transmit such information in electronic form; and (iv) authorizes
the transfer of the Relevant Information to any jurisdiction in which the Relevant Companies consider appropriate. The Grantee shall
have access to, and the right to change, the Relevant Information. Relevant Information will only be used in accordance with applicable
law.
10. Notices.
Notices hereunder shall be mailed or delivered to the Company at its principal place of business and shall be mailed or delivered to
the Grantee at the address on file with the Company or, in either case, at such other address as one party may subsequently furnish to
the other party in writing.
| |
City Therapeutics, Inc. |
| |
|
|
| |
By: |
|
| |
|
Title: |
The foregoing Agreement
is hereby accepted and the terms and conditions thereof hereby agreed to by the undersigned. Electronic acceptance of this Agreement
pursuant to the Company’s instructions to the Grantee (including through an online acceptance process) is acceptable.
| Dated: |
|
|
|
| |
|
|
Grantee’s Signature |
| |
|
|
|
| |
|
|
Grantee’s name and
address: |
| |
|
|
|
| |
|
|
|
| |
|
|
|
| |
|
|
|
| |
|
|
|
| |
|
|
|
Exhibit 10.3
CITY
THERAPEUTICS, INC.
2026
EMPLOYEE STOCK PURCHASE PLAN
The purpose of the City Therapeutics, Inc.
2026 Employee Stock Purchase Plan (the “Plan”) is to provide eligible employees of City Therapeutics, Inc. (the “Company”)
and each Designated Company (as defined in Section 11) with opportunities to purchase shares of the Company’s common stock,
par value $0.0001 per share (the “Common Stock”). 503,322 shares of Common Stock in the aggregate have been approved and reserved
for this purpose, plus on January 1, 2027 and each January 1 thereafter until the Plan terminates pursuant to Section 20,
the number of shares of Common Stock reserved and available for issuance under the Plan shall be cumulatively increased by the least of
(i) 1,509,966 shares of Common Stock, (ii) 1% of the number of Outstanding Shares on the immediately preceding December 31,
and (iii) such number of shares of Common Stock as determined by the Administrator (as defined in Section 1). The Plan includes
two components: a Code Section 423 Component (the “423 Component”) and a non-Code Section 423 Component (the “Non-423
Component”). It is intended for the 423 Component to constitute an “employee stock purchase plan” within the meaning
of Section 423(b) of the U.S. Internal Revenue Code of 1986, as amended (the “Code”), and the 423 Component shall
be interpreted in accordance with that intent. Under the Non-423 Component, which does not qualify as an “employee stock purchase
plan” within the meaning of Section 423(b) of the Code, options will be granted pursuant to rules, procedures or sub-plans
adopted by the Administrator designed to comply with applicable laws or achieve tax and other objectives. Except as otherwise provided
herein or by the Administrator, the Non-423 Component will operate and be administered in the same manner as the 423 Component. Unless
otherwise defined herein, capitalized terms in this Plan shall have the meaning ascribed to them in Section 11.
1. Administration.
The Plan will be administered by the person or persons (the “Administrator”) appointed by the Company’s Board of Directors
(the “Board”) for such purpose. The Administrator has authority at any time to: (i) adopt, alter and repeal such rules,
guidelines and practices for the administration of the Plan and for its own acts and proceedings as it shall deem advisable; (ii) interpret
the terms and provisions of the Plan, including to accommodate the specific requirements of applicable laws, regulations and procedures
in jurisdictions outside of the United States; (iii) make all determinations it deems advisable for the administration of the Plan;
(iv) decide all disputes arising in connection with the Plan; and (v) otherwise supervise the administration of the Plan. All
interpretations and decisions of the Administrator shall be binding on all persons, including the Company and the Participants. No member
of the Board or individual exercising administrative authority with respect to the Plan shall be liable for any action or determination
made in good faith with respect to the Plan or any option granted hereunder.
2. Offerings.
The Company will make one or more offerings to eligible employees to purchase Common Stock under the Plan (“Offerings”) consisting
of one or more Purchase Periods. Each Offering will begin and end on the dates to be determined by the Administrator, provided that no
Offering shall exceed 27 months in duration. Unless the Administrator, in its sole discretion, chooses otherwise prior to an Offering
Date (as defined below), and to the extent an Offering has more than one Purchase Period and to the extent permitted by applicable law,
if the Fair Market Value of the Common Stock on any Exercise Date in an Offering is lower than the Fair Market Value of the Common Stock
on the Offering Date, then all Participants in such Offering automatically will be withdrawn from such Offering immediately after the
exercise of their option on such Exercise Date and automatically re-enrolled in the immediately following Offering as of the first day
thereof and the preceding Offering will terminate.
3. Eligibility.
All individuals classified as employees on the payroll records of the Company and each Designated Company are eligible to participate
in any one or more of the Offerings under the Plan (provided, that the Participant is not permitted to participate in multiple Offerings
at the same time, unless otherwise determined by the Administrator). Notwithstanding the foregoing, an employee will not be eligible to
participate unless, on the first day of the applicable Offering (the “Offering Date”), the employee has been in the employ
of the Company or a Designated Company, as the case may be, for such continuous period preceding such Offering Date as the Administrator
may require, but in no event will the required period of continuous employment be equal to or greater than two years. In addition, the
Administrator may provide that no employee will be eligible to participate under the Plan unless, on the Offering Date, such employee’s
customary employment with the Company or a Designated Company is more than 20 hours per week and more than five months per calendar year
or such other criteria as the Administrator may determine consistent with Section 423 of the Code. Notwithstanding any other provision
herein, individuals who are not contemporaneously classified as employees of the Company or a Designated Company for purposes of the Company’s
or applicable Designated Company’s payroll system are not considered to be eligible employees of the Company or any Designated Company
and shall not be eligible to participate in the Plan. In the event any such individuals are reclassified as employees of the Company or
a Designated Company for any purpose, including, without limitation, common law or statutory employees, by any action of any third party,
including, without limitation, any government agency, or as a result of any private lawsuit, action or administrative proceeding, such
individuals shall, notwithstanding such reclassification, remain ineligible for participation. Notwithstanding the foregoing, the exclusive
means for individuals who are not contemporaneously classified as employees of the Company or a Designated Company on the Company’s
or Designated Company’s payroll system to become eligible to participate in this Plan is through an amendment or subplan to this
Plan, duly executed by the Company, which specifically renders such individuals eligible to participate herein.
4. Participation.
(a) Participants
in Offerings. An eligible employee who is not a Participant in any prior Offering may participate in a subsequent Offering by submitting
an enrollment form to the Company or an agent designated by the Company (in the manner described in Section 4(b) at least 15
business days before the Offering Date (or by such other deadline as shall be established by the Administrator for the Offering).
(b) Enrollment.
The enrollment form (which may be in an electronic format or such other method as determined by the Company in accordance with the Company’s
practices) will (i) state a whole percentage or amount to be deducted from an eligible employee’s Compensation (as defined
in Section 11) per pay period, (ii) authorize the purchase of Common Stock in each Offering in accordance with the terms of
the Plan and (iii) specify the exact name or names in which shares of Common Stock purchased for such individual are to be issued
pursuant to Section 10. An employee who does not enroll in accordance with these procedures will be deemed to have waived the right
to participate. Unless a Participant files a new enrollment form or withdraws from the Plan, such Participant’s deductions or contributions
and purchases will continue at the same percentage or amount of Compensation for future Offerings, provided he or she remains eligible.
(c) Notwithstanding
the foregoing, participation in the Plan will neither be permitted nor be denied contrary to the requirements of the Code.
5. Employee
Contributions. Unless otherwise specified by the Administrator in advance of an Offering, each eligible employee may authorize payroll
deductions or contributions at a minimum of 1 percent (1%) up to a maximum of 15 percent (15%) of such employee’s Compensation
for each pay period. The Company will maintain book accounts showing the amount of payroll deductions or contributions made by each Participant
for each Purchase Period within an Offering. No interest will accrue or be paid on payroll deductions or contributions, except as may
be required by applicable law. If payroll deductions or contributions for purposes of the Plan are prohibited or otherwise problematic
under applicable law (as determined by the Administrator in its discretion), the Administrator may require Participants to contribute
to the Plan by such other means as determined by the Administrator. Any reference to “payroll deductions” or contributions
in this Section 5 (or in any other section of the Plan) will similarly cover contributions by other means made pursuant to this Section 5.
6. Deduction
Changes. Except as may be determined by the Administrator in advance of an Offering, a Participant may not increase or decrease his
or her payroll deduction or contributions during any Offering, but may increase or decrease his or her payroll deduction or contributions
with respect to the next Offering (subject to the limitations of Section 5) by filing a new enrollment form at least fifteen (15)
business days before the next Offering Date (or by such other deadline as shall be established by the Administrator for the Offering).
The Administrator may, in advance of any Offering, establish rules permitting a Participant to increase, decrease or terminate his
or her payroll deduction or contributions during an Offering.
7. Withdrawal.
A Participant may withdraw from participation in the Plan by delivering a written notice of withdrawal to the Company or an agent designated
by the Company (in accordance with such procedures as may be established by the Administrator). The Participant’s withdrawal will
be effective as of the next business day (or such other date established by the Administrator). Following a Participant’s withdrawal,
the Company will promptly refund such individual’s entire account balance under the Plan to him or her (after payment for any Common
Stock purchased before the effective date of withdrawal). Partial withdrawals are not permitted. Such an employee may not begin participation
again during the remainder of the Offering, but may enroll in a subsequent Offering in accordance with Section 4.
8. Grant
of Options. On each Offering Date, the Company will grant to each eligible employee who is then a Participant in the Plan an option
(“Option”) to purchase, on the last day of a Purchase Period (an “Exercise Date”) and at the Option Price (as
defined herein) hereinafter provided for, the lower of (a) a number of shares of Common Stock determined by dividing such Participant’s
accumulated payroll deductions or contributions on such Exercise Date by the Option Price or (b) a number of shares of Common Stock
determined by dividing $25,000 by the Fair Market Value of the Common Stock on the Offering Date for such Offering (or such other maximum
number of shares as shall have been established by the Administrator in advance of the Offering); provided, however, that such Option
shall be subject to the limitations set forth below. Each Participant’s Option shall be exercisable only to the extent of such Participant’s
accumulated payroll deductions or contributions on the Exercise Date. The purchase price for each share purchased under each Option (the
“Option Price”) will be 85 percent (85%) of the Fair Market Value (as defined in Section 11) of the Common Stock on the
Offering Date or the Exercise Date, whichever is less.
Notwithstanding the foregoing,
no Participant may be granted an Option hereunder if such Participant, immediately after the Option was granted, would be treated as owning
stock possessing 5 percent (5%) or more of the total combined voting power or value of all classes of stock of the Company or any
Parent or Subsidiary (as defined in Section 11). For purposes of the preceding sentence, the attribution rules of Section 424(d) of
the Code shall apply in determining the stock ownership of a Participant, and all stock which the Participant has a contractual right
to purchase shall be treated as stock owned by the Participant. In addition, no Participant may be granted an Option which permits his
or her rights to purchase stock under the Plan, and any other employee stock purchase plan of the Company and its Parents and Subsidiaries,
to accrue at a rate which exceeds $25,000 of the Fair Market Value of the Common Stock (determined on the option grant date or dates)
for each calendar year in which the Option is outstanding at any time. The purpose of the limitation in the preceding sentence is to comply
with Section 423(b)(8) of the Code and shall be applied taking Options into account in the order in which they were granted.
9. Exercise
of Option and Purchase of Shares. Each employee who continues to be a Participant in the Plan on an Exercise Date shall be deemed
to have exercised his or her Option on such date and shall acquire from the Company such number of whole shares of Common Stock reserved
for the purpose of the Plan as his or her accumulated payroll deductions on such date will purchase at the Option Price, subject to any
other limitations contained in the Plan. Unless otherwise determined by the Administrator in advance of an Offering, any amount remaining
in a Participant’s account after the purchase of shares on an Exercise Date of an Offering solely by reason of the inability to
purchase a fractional share will be carried forward to the next Purchase Period; provided, that if such Exercise Date is the final Exercise
Date of an Offering, such amount will be carried forward to the next Offering and any other balance remaining in a Participant’s
account at the end of an Offering will be refunded to the Participant promptly.
10. Issuance
of Certificates. Certificates or book-entries at the Company’s transfer agent representing shares of Common Stock purchased
under the Plan may be issued only in the name of the employee, in the name of the employee and another person of legal age as joint tenants
with rights of survivorship or in the name of a broker authorized by the employee to be his, her or their nominee for such purpose.
11. Definitions.
The term “Affiliate”
means any entity that, directly or indirectly through one or more intermediaries, controls, is controlled by or is under the common control
with, the Company.
The term “Compensation”
means the amount of base pay, prior to salary reduction pursuant to Sections 125, 132(f) or 401(k) of the Code, but excluding
overtime, commissions, incentive or bonus awards, allowances and reimbursements for expenses such as relocation allowances or travel expenses,
income or gains related to Company stock options or other share-based awards and similar items. The Administrator shall have the discretion
to (i) determine the application of this definition to Participants outside the United States and (ii) on a uniform and nondiscriminatory
basis, establish a different definition of Compensation in advance of any Offering (and, with respect to the 423 Component, subject to
compliance of such definition with Section 423 of the Code).
The term “Designated
Company” means any present or future Subsidiary or Affiliate that has been designated by the Administrator to participate in the
Plan. The Administrator may designate any Subsidiary or Affiliate, or revoke any such designation, at any time and from time to time,
either before or after the Plan is approved by the stockholders, and may further designate such companies or Participants as participating
in the 423 Component or the Non-423 Component. The Administrator may also determine which Affiliates or eligible employees may be excluded
from participation in the Plan, to the extent consistent with Section 423 of the Code or as implemented under the Non-423 Component,
and determine which Designated Company or Companies will participate in separate Offerings (to the extent that the Company makes separate
Offerings). For purposes of the 423 Component, only the Company and its Subsidiaries may be Designated Companies; provided, however, that
at any given time, a Subsidiary that is a Designated Company under the 423 Component will not be a Designated Company under the Non-423
Component.
The term “Fair Market
Value of the Common Stock” on any given date means the fair market value of the Common Stock determined in good faith by the Administrator;
provided, however, that if the Common Stock is admitted to quotation on the National Association of Securities Dealers Automated Quotation
System (“Nasdaq”), Nasdaq Global Market, The New York Stock Exchange or another national securities exchange or traded on
any established market, the determination shall be made by reference to the closing price on such date. If there is no closing price for
such date, the determination shall be made by reference to the last date preceding such date for which there is a closing price.
The term “Initial Public
Offering” means the first underwritten, firm commitment public offering pursuant to an effective registration statement under the
U.S. Securities Act of 1933, as amended, covering the offer and sale by the Company of its Common Stock.
The term “New Exercise
Date” means a new Exercise Date if the Administrator shortens any Offering then in progress.
The term “Outstanding
Shares” means, as of a specified date, the sum of (a) the number of shares of Stock issued and outstanding and (b) the
number of shares of Stock issuable pursuant to the exercise of any outstanding, pre-funded warrants to acquire Stock for a nominal exercise
price.
The term “Parent”
means a “parent corporation” with respect to the Company, as defined in Section 424(e) of the Code.
The term “Participant”
means an individual who is eligible as determined in Section 3 and who has complied with the provisions of Section 4.
The term “Purchase Period”
means a period of time specified within an Offering, as may be specified by the Administrator in accordance with Section 2, generally
beginning on the Offering Date or the next day following an Exercise Date within an Offering, and ending on an Exercise Date. An Offering
may consist of one or more Purchase Periods.
The term “Registration
Date” means the date on which the registration statement on Form S-1 is filed by the Company with respect to its Initial Public
Offering is declared effective by the U.S. Securities and Exchange Commission (the “SEC”).
The
term “Sale Event” means (i) the sale of all or substantially all of the assets of the Company on a consolidated basis
to an unrelated person or entity, (ii) a merger, reorganization, statutory share exchange, consolidation, or similar transaction
pursuant to which the holders of the Company’s outstanding voting power and outstanding stock immediately prior to such transaction
do not own a majority of the outstanding voting power and outstanding stock or other equity interests of the resulting or successor entity
(or its ultimate parent, if applicable) immediately upon completion of such transaction, (iii) the sale of all of the Common Stock
to an unrelated person, entity or group thereof acting in concert, (iv) any other transaction in which the owners of the Company’s
outstanding voting power immediately prior to such transaction do not own at least a majority of the outstanding voting power of the Company
or any successor entity immediately upon completion of the transaction other than as a result of the acquisition of securities directly
from the Company, or (v) the approval by the stockholders of the Company of a complete liquidation or dissolution of the Company.
The term “Subsidiary” means a “subsidiary
corporation” with respect to the Company, as defined in Section 424(f) of the Code.
12. Rights
on Termination or Transfer of Employment. If a Participant’s employment terminates for any reason before the Exercise Date for
any Offering, no payroll deduction or contributions will be taken from any pay due and owing to the Participant and the balance in the
Participant’s account will be paid to such Participant or, in the case of such Participant’s death, if permitted by the Administrator
and valid under applicable law, to his or her designated beneficiary or to the legal representative of his or her estate as if such Participant
had withdrawn from the Plan under Section 7. An employee will be deemed to have terminated employment, for this purpose, if the corporation
that employs him or her, having been a Designated Company, ceases to be a Subsidiary or Affiliate, or if the employee is transferred to
any corporation other than the Company or a Designated Company. Unless otherwise determined by the Administrator, a Participant whose
employment transfers between, or whose employment terminates with an immediate rehire (with no break in service) by, Designated Companies
or a Designated Company and the Company, will not be treated as having terminated employment for purposes of participating in the Plan
or an Offering; provided, however, that if a Participant transfers from an Offering under the 423 Component to an Offering under the Non-423
Component, the exercise of the Participant’s Option will be qualified under the 423 Component only to the extent that such exercise
complies with Section 423 of the Code. If a Participant transfers from an Offering under the Non-423 Component to an Offering under
the 423 Component, the exercise of the Participant’s Option will remain non-qualified under the Non-423 Component. Further, an employee
will not be deemed to have terminated employment for purposes of this Section 12 if the employee is on an approved leave of absence
for military service or sickness or for any other purpose approved by the Company, if the employee’s right to reemployment is guaranteed
either by a statute or by contract or under the policy pursuant to which the leave of absence was granted or if the Administrator otherwise
provides in writing.
13. Special
Rules and Sub-Plans. Notwithstanding anything herein to the contrary, the Administrator may adopt special rules or sub-plans
applicable to the employees of a particular Designated Company, whenever the Administrator determines that such rules are necessary
or appropriate for the implementation of the Plan in a jurisdiction where such Designated Company has employees, regarding, without limitation,
eligibility to participate in the Plan, handling and making of payroll deductions or contributions by other means, establishment of bank
or trust accounts to hold payroll deductions or contributions, payment of interest, conversion of local currency, obligation to pay payroll
tax, withholding procedures and handling of share issuances, any of which may vary according to applicable requirements; provided that
if such special rules or sub-plans are inconsistent with the requirements of Section 423(b) of the Code, the employees
subject to such special rules or sub-plans will participate in the Non-423 Component.
14. Optionees
Not Stockholders. Neither the granting of an Option to a Participant nor the deductions or contributions from his or her pay shall
result in such Participant becoming a holder of the shares of Common Stock covered by an Option under the Plan until such shares have
been purchased by and issued to him or her.
15. Rights
Not Transferable. Rights under the Plan are not transferable by a Participant other than by will or the laws of descent and distribution,
and are exercisable during the Participant’s lifetime only by the Participant.
16. Application
of Funds. All funds received or held by the Company under the Plan may be combined with other corporate funds and may be used for
any corporate purpose, unless otherwise required under applicable law.
17. Adjustment
in Case of Changes Affecting Common Stock. In the event of a subdivision of outstanding shares of Common Stock, the payment of a dividend
in Common Stock or any other change affecting the Common Stock, the number of shares approved for the Plan and the share limitation set
forth in Section 8 shall be equitably or proportionately adjusted to give proper effect to such event. In the case of and subject
to the consummation of a Sale Event, the Administrator, in its discretion, on such terms and conditions as it deems appropriate, is hereby
authorized to take any one or more of the following actions whenever the Administrator determines that such action is appropriate in order
to prevent the dilution or enlargement of the benefits or potential benefits intended to be made available under the Plan or with respect
to any right under the Plan or to facilitate such transactions or events:
(a) To
provide for either (i) termination of any outstanding Option in exchange for an amount of cash, if any, equal to the amount that
would have been obtained upon the exercise of such Option had such Option been currently exercisable or (ii) the replacement of such
outstanding Option with other options or property selected by the Administrator in its sole discretion.
(b) To
provide that the outstanding Options under the Plan shall be assumed by the successor or survivor corporation, or a parent or subsidiary
thereof, or shall be substituted for similar options covering the stock of the successor or survivor corporation, or a parent or subsidiary
thereof, with appropriate adjustments as to the number and kind of shares and prices.
(c) To
make adjustments in the number and type of shares of Common Stock (or other securities or property) subject to outstanding Options under
the Plan and/or in the terms and conditions of outstanding Options and Options that may be granted in the future.
(d) To
provide that the Offering with respect to which an Option relates will be shortened by setting a New Exercise Date on which such Offering
will end. The New Exercise Date will occur before the date of the Sale Event. The Administrator will notify each Participant in writing
or electronically prior to the New Exercise Date, that the Exercise Date for the Participant’s Option has been changed to the New
Exercise Date and that the Participant’s Option will be exercised automatically on the New Exercise Date, unless prior to such date
the Participant has withdrawn from the Offering as provided in Section 7 hereof.
(e) To
provide that all outstanding Options shall terminate without being exercised and all amounts in the accounts of Participants shall be
promptly refunded.
18. Amendment
of the Plan. The Board may at any time and from time to time amend the Plan in any respect, except that without the approval within
12 months of such Board action by the stockholders, no amendment shall be made increasing the number of shares approved for the Plan or
making any other change that would require stockholder approval in order for the Section 423 Component of the Plan, as amended, to
qualify as an “employee stock purchase plan” under Section 423(b) of the Code.
19. Insufficient
Shares. If the total number of shares of Common Stock that would otherwise be purchased on any Exercise Date plus the number of shares
purchased under previous Offerings under the Plan exceeds the maximum number of shares issuable under the Plan, the shares then available
shall be apportioned among Participants in proportion to the amount of payroll deductions or contributions accumulated on behalf of each
Participant that would otherwise be used to purchase Common Stock on such Exercise Date.
20. Termination
of the Plan. The Plan may be terminated at any time by the Board. Upon termination of the Plan, all amounts in the accounts of Participants
shall be promptly refunded. Unless terminated earlier, the Plan shall expire on the ten-year anniversary of the Registration Date.
21. Governmental
Regulations. The Company’s obligation to sell and deliver Common Stock under the Plan is subject to applicable laws and the
completion of any registration or qualification of the Common Stock under any U.S. or non-U.S., local, state or federal securities or
exchange control law, or under rulings or regulations of the SEC or of any other governmental regulatory body, and to obtaining any approval
or other clearance from any U.S. or non-U.S. local, state or federal governmental agency, which registration, qualification or approval
the Company shall, in its absolute discretion, deem necessary or advisable. The Company is under no obligation to register or qualify
the Common Stock with the SEC or any other U.S. or non-U.S. securities commission or to seek approval or clearance from any governmental
authority for the issuance or sale of such stock.
22. Governing
Law. This Plan and all Options and actions taken thereunder shall be governed by, and construed in accordance with, the General Corporation
Law of the State of Delaware as to matters within the scope thereof, and as to all other matters shall be governed and construed in accordance
with the internal laws of the State of Delaware, applied without regard to conflict of law principles.
23. Issuance
of Shares. Shares may be issued upon exercise of an Option from authorized but unissued Common Stock, from shares held in the treasury
of the Company or from any other proper source.
24. Tax
Withholding. Participation in the Plan is subject to any applicable U.S. and non-U.S. federal, state or local tax withholding requirements
on income the Participant realizes in connection with the Plan. Each Participant agrees, by entering the Plan, that the Company or any
Subsidiary or Affiliate may withhold from a Participant’s wages, salary or other compensation at any time the amount necessary for
the Company or any Subsidiary or Affiliate to meet applicable withholding obligations, including any withholding required to make available
to the Company or any Subsidiary or Affiliate any tax deductions or benefits attributable to the sale or disposition of Common Stock by
such Participant. In addition, the Company or any Subsidiary or Affiliate may withhold from the proceeds of the sale of Common Stock or
use any other method of withholding that the Company or any Subsidiary or Affiliate deems appropriate to the extent permitted by U.S.
Treasury Regulation Section 1.423-2(f) with respect to the 423 Component. The Company will not be required to issue any Common
Stock under the Plan until such obligations are satisfied.
25. Notification
Upon Sale of Shares Under the 423 Component. Each Participant agrees, by entering the 423 Component of the Plan, to give the Company
prompt notice of any disposition of shares purchased under the Plan where such disposition occurs within two years after the date of grant
of the Option pursuant to which such shares were purchased or within one year after the date such shares were purchased.
26. Effective
Date and Approval of Shareholders. The Plan shall take effect on the date immediately preceding the Registration Date, subject to
approval by the holders of a majority of the votes cast at a meeting of stockholders at which a quorum is present or by written consent
of the stockholders (the “Effective Date”).
27. Equal
Rights and Privileges. Notwithstanding any provision of the Plan to the contrary and in accordance with Section 423 of the Code
for the 423 Component of the Plan, all eligible employees who are granted Options under the Plan shall have the same rights and privileges.
28. No
Right to Continued Service. Neither the Plan nor any compensation paid hereunder will confer on any Participant the right to continue
as an employee or in any other capacity.
29. Entire
Plan. This Plan constitutes the entire plan with respect to the subject matter hereof and supersedes all prior plans with respect
to the subject matter hereof.
DATE APPROVED BY BOARD OF DIRECTORS: September 19, 2026
DATE APPROVED BY STOCKHOLDERS: October 8, 2026
Exhibit 10.6
City
Therapeutics, Inc.
SENIOR EXECUTIVE CASH INCENTIVE BONUS PLAN
This Senior Executive Cash
Incentive Bonus Plan (the “Incentive Plan”) is intended to provide an incentive for superior work and to motivate eligible
executives of City Therapeutics, Inc. (the “Company”) and its subsidiaries toward even higher achievement and business
results, to tie their goals and interests to those of the Company and its stockholders and to enable the Company to attract and retain
highly qualified executives. The Incentive Plan is for the benefit of Covered Executives (as defined below).
From time to time, the Culture
and Compensation Committee of the Board of Directors (the “Board”) of the Company (the “Culture and Compensation Committee”)
may select certain key executives (the “Covered Executives”) to be eligible to receive bonuses hereunder. Participation in
the Incentive Plan does not change the “at will” nature of a Covered Executive’s employment with the Company.
The Culture and Compensation
Committee shall have the sole discretion and authority to administer and interpret the Incentive Plan.
(a) Corporate
Performance Goals. A Covered Executive may receive a bonus payment under the Incentive Plan based upon the attainment of one or more
performance objectives that are established by the Culture and Compensation Committee, together with management, and approved by the Board
and relate to financial and operational metrics with respect to the Company or any of its subsidiaries (the “Corporate Performance
Goals”), including, but not limited to: research and development, publication, clinical and/or regulatory milestones; cash flow
(including, but not limited to, operating cash flow and free cash flow); revenue; earnings before interest, taxes, depreciation and amortization;
net income (loss) (either before or after interest, taxes, depreciation and/or amortization); changes in the market price of the Company’s
common stock; economic value-added; acquisitions, licenses, collaborations or strategic transactions, including government or other third-party
grants, asset sales, royalty financings, partnership, collaboration and licensing arrangements, or a combination of these approaches;
financing or other capital raising transactions; operating income (loss); return on capital, assets, equity, or investment; stockholder
returns; return on sales; gross or net profit levels; productivity; expense efficiency; margins; operating efficiency;; working capital;
earnings (loss) per share of the Company’s common stock; sales or market shares; number of prescriptions or prescribing physicians;
coverage decisions; leadership development, employee retention and recruiting, and other human resources matters; operating income and/or
net annual recurring revenue, or any other performance goal selected by the Culture and Compensation Committee, any of which may be (A) measured
in absolute terms or compared to any incremental increase, (B) measured in terms of growth, (C) compared to another company
or companies or to results of a peer group, (D) measured against the market as a whole and/or as compared to applicable market indices
and/or (E) measured on a pre-tax or post-tax basis (if applicable). Further, any Corporate Performance Goals may be used to measure
the performance of the Company as a whole or a business unit or other segment of the Company, or one or more product lines or specific
markets. The Corporate Performance Goals apply to each Covered Executive for the applicable performance period.
(b) Calculation
of Corporate Performance Goals. At the beginning of each applicable performance period, the Culture and Compensation Committee will
determine whether any significant element(s) will be included in or excluded from the calculation of any Corporate Performance Goal
with respect to the Covered Executives. In all other respects, Corporate Performance Goals will be calculated in accordance with
the Company’s financial statements, generally accepted accounting principles or under a methodology established by the Culture and
Compensation Committee at the beginning of the performance period and which is consistently applied with respect to a Corporate Performance
Goal in the relevant performance period.
(c) Target;
Minimum; Maximum. Each Corporate Performance Goal shall have a “target” (i.e., 100 percent attainment of the Corporate
Performance Goal) and may also have a “minimum” hurdle and/or a “maximum” amount.
(d) Bonus
Requirements; Individual Goals. Except as otherwise set forth in this Section 4(d): (i) any bonuses paid to Covered Executives
under the Incentive Plan shall be based upon objectively determinable bonus formulas that tie such bonuses to one or more performance
targets relating to the Corporate Performance Goals, (ii) bonus formulas for Covered Executives shall be adopted in each performance
period by the Culture and Compensation Committee and communicated to each Covered Executive at the beginning of each performance period
and (iii) no bonuses shall be paid to Covered Executives unless and until the Board makes a determination with respect to the attainment
of the performance targets relating to the Corporate Performance Goals based upon the recommendation of the Culture and Compensation Committee.
Notwithstanding the foregoing, the Culture and Compensation Committee may adjust bonuses payable under the Incentive Plan based on achievement
of one or more individual performance objectives or pay bonuses (including, without limitation, discretionary bonuses) to Covered Executives
under the Incentive Plan based on individual performance goals and/or upon such other terms and conditions as the Culture and Compensation
Committee may in its discretion determine.
(e) Individual
Target Bonuses. The Culture and Compensation Committee shall establish a target bonus opportunity for each Covered Executive for each
performance period. For each Covered Executive, the Culture and Compensation Committee shall have the authority to apportion the target
award so that a portion of the target award shall be tied to attainment of Corporate Performance Goals and a portion of the target award
shall be tied to attainment of individual performance objectives.
(f) Employment
Requirement. Subject to any additional terms contained in a written agreement between the Covered Executive and the Company, the payment
of a bonus to a Covered Executive with respect to a performance period shall be conditioned upon the Covered Executive’s employment
by the Company on the bonus payment date. If a Covered Executive was not employed for an entire performance period, the Culture and Compensation
Committee may pro rate the bonus based on the number of days employed during such period.
(a) With
respect to Corporate Performance Goals established and measured on an annual basis, Corporate Performance Goals will be measured as of
the end of the fiscal year after the Company’s financial reports with respect to such period(s) have been published, as applicable.
If the Corporate Performance Goals for any such period are met, bonus payments will be made as soon as practicable, but not later than
two and one-half months after the end of the fiscal year in which the applicable performance period ends.
(b) For
the avoidance of doubt, bonuses earned at any time in a fiscal year must be paid no later than two and one-half months after the last
day of such fiscal year.
| 6. | Amendment and Termination |
The Company reserves the right
to amend or terminate the Incentive Plan at any time in its sole discretion.
| 7. | Company Recoupment Rights |
A Covered Executive’s
rights with respect to any award granted pursuant to the Incentive Plan shall in all events be subject to reduction, cancellation, forfeiture
or recoupment to the extent necessary to comply with (i) any right that the Company may have under any Company clawback, forfeiture
or recoupment policy as in effect from time to time or other agreement or arrangement with a Covered Executive, or (ii) applicable
law.
Adopted September 19, 2026, subject to effectiveness
of the Company’s Registration Statement on Form S-1.
Exhibit 10.7
City
Therapeutics, Inc.
NON-EMPLOYEE DIRECTOR COMPENSATION POLICY
The purpose of this
Non-Employee Director Compensation Policy (the “Policy”) of City Therapeutics, Inc. (the “Company”) is to
provide a total compensation package that enables the Company to attract and retain, on a long-term basis, high-caliber directors who
are not employees or officers of the Company or its subsidiaries (“Outside Directors”). This Policy will become effective
as of the effective time of the registration statement for the Company’s initial public offering of its equity securities (the “Effective
Date”). Unless otherwise determined by the Board of Directors, Outside Directors who are employees of, or otherwise affiliated
with, an institutional investor in the Company shall not be eligible to receive compensation under this Policy. In furtherance of the
purpose stated above, all Outside Directors shall be paid compensation for services provided to the Company as Outside Directors as set
forth below:
Cash Retainers
Annual Retainer
for Board Membership: $40,000 for general availability and participation in meetings and conference calls of our Board of Directors,
to be paid quarterly in arrears, pro-rated based on the number of actual days served by the director during such calendar quarter. No
additional compensation will be paid for attending individual meetings of the Board of Directors.
| Additional Annual Retainer for Non-Executive
Chairperson: | |
$ | 30,000 | |
| | |
| | |
| Additional Annual Retainers for Committee Membership: | |
| | |
| | |
| | |
| Audit Committee Chairperson: | |
$ | 20,000 | |
| | |
| | |
| Audit Committee member (other than Chairperson): | |
$ | 10,000 | |
| | |
| | |
| Compensation Committee Chairperson: | |
$ | 15,000 | |
| | |
| | |
| Compensation Committee member (other than Chairperson): | |
$ | 7,500 | |
| | |
| | |
| Nominating and Corporate Governance Committee Chairperson: | |
$ | 10,000 | |
| | |
| | |
| Nominating and Corporate Governance Committee member (other than Chairperson): | |
$ | 5,000 | |
Equity Retainers
All grants of equity retainer awards to Outside Directors pursuant
to this Policy will be made in accordance with the following provisions:
IPO Award:
Upon the Effective Date, each Outside Director as of such date will receive an Annual Award (as defined below) (the “IPO Award”).
IPO Awards will have an exercise price per share equal to the per share “price to the public” (or equivalent) set forth on
the cover page for the final prospectus relating to the Company’s initial public offering, expire ten years from the date of
grant and vest in full on the earlier of (i) the first anniversary of the date of grant or (ii) the date of the next Annual
Meeting following the grant date; provided, however, that all vesting shall cease if the director resigns from the Board of Directors
or otherwise ceases to serve on the Board of Directors of the Company, unless the Board of Directors determines that the circumstances
warrant continuation of vesting.
Initial Award:
Upon his or her initial election or appointment to the Board of Directors, each Outside Director will receive an initial, one-time stock
option award (the “Initial Award”) to purchase 45,802 shares, which shall vest in equal monthly installments over three years
from the date of grant, provided, however, that all vesting shall cease if the director resigns from the Board of Directors or otherwise
ceases to serve on the Board of Directors of the Company, unless the Board of Directors determines that the circumstances warrant continuation
of vesting. The Initial Award shall expire ten years from the date of grant, and shall have a per share exercise price equal to the Fair
Market Value (as defined in the Company’s 2026 Stock Option and Incentive Plan) of the Company’s common stock on the date
of grant. This Initial Award applies only to Outside Directors who are first elected to the Board of Directors subsequent to the Effective
Date.
Annual Award:
On each date of each Annual Meeting of Stockholders of the Company following the Effective Date (the “Annual Meeting”), each
continuing Outside Director, other than a director receiving an Initial Award, will receive an annual stock option award (the “Annual
Award”) to purchase 22,649 shares, which shall vest in full upon the earlier of (i) the first anniversary of the date of grant
or (ii) the date of the next Annual Meeting; provided, however, that all vesting shall cease if the director resigns from the Board
of Directors or otherwise ceases to serve on the Board of Directors of the Company, unless the Board of Directors determines that the
circumstances warrant continuation of vesting. The Annual Award shall expire ten years from the date of grant, and shall have a per share
exercise price equal to the Fair Market Value of the Company’s common stock on the date of grant.
Sale Event Acceleration:
All outstanding Initial Awards and Annual Awards held by an Outside Director shall become fully vested and exercisable upon a Sale Event
(as defined in the Company’s 2026 Stock Option and Incentive Plan).
Expenses
The Company will reimburse all reasonable out-of-pocket
expenses incurred by Outside Directors in attending meetings of the Board of Directors or any committee thereof.
Maximum Annual Compensation
The aggregate amount of compensation, including
both equity compensation and cash compensation, paid by the Company to any Outside Director in a calendar year for services as an Outside
Director shall not exceed $750,000 (or such other limit as may be set forth in Section 3(b) of the Company’s 2026 Stock
Option and Incentive Plan, as amended from time to time, or any similar provision of a successor plan); provided, however, that in the
first calendar year in which an individual becomes an Outside Director, the aggregate amount of all equity compensation awarded and all
other cash compensation paid by the Company to such Outside Director for services as an Outside Director shall not exceed $1,000,000 (or
such other limit as may be set forth in Section 3(b) of the Company’s 2026 Stock Option and Incentive Plan, as amended
from time to time, or any similar provision of a successor plan). For this purpose, the “amount” of equity compensation paid
in a calendar year shall be determined based on the grant date fair value thereof, as determined in accordance with FASB ASC Topic 718
or its successor provision, but excluding the impact of estimated forfeitures related to service-based vesting conditions.
Adopted September 19, 2026, subject to effectiveness of the Company’s
Registration Statement on Form S-1.
Exhibit 10.8
City
Therapeutics, Inc.
COMPENSATION RECOVERY POLICY
Adopted as of September 19, 2026, subject
to effectiveness of the Company’s Registration Statement on Form S-1 for its initial public offering.
City Therapeutics, Inc.,
a Delaware corporation (the “Company”), has adopted a Compensation Recovery Policy (this “Policy”) as described
below.
The Policy sets forth the circumstances and procedures
under which the Company shall recover Erroneously Awarded Compensation from Covered Persons (as defined below) in accordance with rules issued
by the United States Securities and Exchange Commission (the “SEC”) under the Securities Exchange Act of 1934, as amended
(the “Exchange Act”), and the Nasdaq Global Market. Capitalized terms used and not otherwise defined herein shall have the
meanings given in Section 3 below.
| 2. | Compensation Recovery Requirement |
In the event the Company is required to prepare
a Financial Restatement, the Company shall recover reasonably promptly all Erroneously Awarded Compensation with respect to such Financial
Restatement.
| a. | “Applicable Recovery Period” means the three completed fiscal years immediately preceding
the Restatement Date for a Financial Restatement. In addition, in the event the Company has changed its fiscal year: (i) any transition
period of less than nine months occurring within or immediately following such three completed fiscal years shall also be part of such
Applicable Recovery Period and (ii) any transition period of nine to 12 months will be deemed to be a completed fiscal year. |
| b. | “Applicable Rules” means any rules or regulations adopted by the Exchange pursuant to
Rule 10D-1 under the Exchange Act and any applicable rules or regulations adopted by the SEC pursuant to Section 10D of
the Exchange Act. |
| c. | “Board” means the Board of Directors of the Company. |
| d. | “Committee” means the Compensation Committee of the Board or, in the absence of such committee,
a majority of independent directors serving on the Board. |
| e. | “Covered Person” means any Executive Officer. A person’s status as a Covered Person
with respect to Erroneously Awarded Compensation shall be determined as of the time of receipt of such Erroneously Awarded Compensation
regardless of the person’s current role or status with the Company (e.g., if a person began service as an Executive Officer after
the beginning of an Applicable Recovery Period, that person would not be considered a Covered Person with respect to Erroneously Awarded
Compensation received before the person began service as an Executive Officer, but would be considered a Covered Person with respect to
Erroneously Awarded Compensation received after the person began service as an Executive Officer where such person served as an Executive
Officer at any time during the performance period for such Erroneously Awarded Compensation). |
| f. | “Effective Date” means the date of effectiveness of the Company’s Registration Statement
on Form S-1 for its initial public offering. |
| g. | “Erroneously Awarded Compensation” means the amount of any Incentive-Based Compensation received
by a Covered Person on or after the Effective Date and during the Applicable Recovery Period that exceeds the amount that otherwise would
have been received by the Covered Person had such compensation been determined based on the restated amounts in a Financial Restatement,
computed without regard to any taxes paid. Calculation of Erroneously Awarded Compensation with respect to Incentive-Based Compensation
based on stock price or total shareholder return, where the amount of Erroneously Awarded Compensation is not subject to mathematical
recalculation directly from the information in a Financial Restatement, shall be based on a reasonable estimate of the effect of the Financial
Restatement on the stock price or total shareholder return upon which the Incentive-Based Compensation was received, and the Company shall
maintain documentation of the determination of such reasonable estimate and provide such documentation to the Exchange in accordance with
the Applicable Rules. Incentive-Based Compensation is deemed received, earned or vested when the Financial Reporting Measure is attained,
not when the actual payment, grant or vesting occurs. |
| h. | “Exchange” means the Nasdaq Global Market. |
| i. | An “Executive Officer” means any person who served the Company in any of the following roles
at any time during the performance period applicable to Incentive-Based Compensation and received Incentive-Based Compensation after beginning
service in any such role (regardless of whether such Incentive-Based Compensation was received during or after such person’s service
in such role): the president, principal financial officer, principal accounting officer (or if there is no such accounting officer the
controller), any vice president in charge of a principal business unit, division or function (such as sales, administration or finance),
any other officer who performs a policy making function or any other person who performs similar policy making functions for the Company.
Executive officers of parents or subsidiaries of the Company may be deemed executive officers of the Company if they perform such policy
making functions for the Company. |
| j. | “Financial Reporting Measures” mean measures that are determined and presented in accordance
with the accounting principles used in preparing the Company’s financial statements, any measures that are derived wholly or in
part from such measures (including, for example, a non-GAAP financial measure), and stock price and total shareholder return. |
| k. | A “Financial Restatement” means a restatement of previously issued financial statements of
the Company due to the material noncompliance of the Company with any financial reporting requirement under the securities laws, including
any required restatement to correct an error in previously-issued financial statements that is material to the previously-issued financial
statements or that would result in a material misstatement if the error were corrected in the current period or left uncorrected in the
current period. |
| l. | “Incentive-Based Compensation” means any compensation provided, directly or indirectly, by
the Company or any of its subsidiaries that is granted, earned or vested based, in whole or in part, upon the attainment of a Financial
Reporting Measure. For avoidance of doubt, Incentive-Based Compensation is “received” for purposes of this Policy in
the fiscal period during which the Financial Reporting Measure specified in the Incentive-Based Compensation award is attained, even if
the payment or grant of such Incentive-Based Compensation occurs after the end of that period. |
| m. | “Restatement Date” means, with respect to a Financial Restatement, the earlier to occur of:
(i) the date the Board concludes, or reasonably should have concluded, that the Company is required to prepare the Financial Restatement
or (ii) the date a court, regulator or other legally authorized body directs the Company to prepare the Financial Restatement. |
| 4. | Exception to Compensation Recovery Requirement |
The Company may elect not to recover Erroneously
Awarded Compensation pursuant to this Policy if the Committee determines that recovery would be impracticable, and one or more of the
following conditions, together with any further requirements set forth in the Applicable Rules, are met: (i) the direct expense paid
to a third party, including outside legal counsel, to assist in enforcing this Policy would exceed the amount to be recovered, and the
Company has made a reasonable attempt to recover such Erroneously Awarded Compensation; or (ii) recovery would likely cause an otherwise
tax-qualified retirement plan to fail to be so qualified under applicable regulations.
To the extent that, pursuant to this Policy, the
Company is entitled to recover any Erroneously Awarded Compensation that is received by a Covered Person, the gross amount received (i.e.,
the amount the Covered Person received, or was entitled to receive, before any deductions for tax withholding or other payments) shall
be returned by the Covered Person.
| 6. | Method of Compensation Recovery |
The Committee shall determine, in its sole discretion,
the method for recovering Erroneously Awarded Compensation hereunder, which may include, without limitation, any one or more of the following:
| a. | requiring reimbursement of cash Incentive-Based Compensation previously paid; |
| b. | seeking recovery of any gain realized on the vesting, exercise, settlement, sale, transfer or other disposition
of any equity-based awards; |
| c. | cancelling or rescinding some or all outstanding vested or unvested equity-based awards; |
| d. | adjusting or withholding from unpaid compensation or other set-off; |
| e. | cancelling or offsetting against planned future grants of equity-based awards; and/or |
| f. | any other method permitted by applicable law or contract. |
The Committee need not utilize the same method
of recovery for all Covered Persons or with respect to all types of Erroneously Awarded Compensation.
Notwithstanding the foregoing, a Covered Person
will be deemed to have satisfied such person’s obligation to return Erroneously Awarded Compensation to the Company if such Erroneously
Awarded Compensation is returned in the exact same form in which it was received; provided that equity withheld to satisfy tax obligations
will be deemed to have been received in cash in an amount equal to the tax withholding payment made.
In the event the Company is required to recover
Erroneously Awarded Compensation from a Covered Person who is no longer an employee, the Company is entitled to seek such recovery in
order to comply with applicable law, regardless of the terms of any release of claims or separation agreement such individual may have
signed.
This Policy shall be interpreted in a manner that
is consistent with the Applicable Rules and any other applicable law. The Committee shall take into consideration any applicable
interpretations and guidance of the SEC in interpreting this Policy, including, for example, in determining whether a financial restatement
qualifies as a Financial Restatement hereunder. To the extent the Applicable Rules require recovery of Incentive-Based Compensation
in additional circumstances besides those specified above, nothing in this Policy shall be deemed to limit or restrict the right or obligation
of the Company to recover Incentive-Based Compensation to the fullest extent required by the Applicable Rules.
This Policy shall be administered by the Committee;
provided, however, that the Board shall have exclusive authority to authorize the Company to prepare a Financial Restatement. In doing
so, the Board may rely on a recommendation of the Audit Committee of the Board. The Committee shall have such powers and authorities related
to the administration of this Policy as are consistent with the governing documents of the Company and applicable law. The Committee shall
have full power and authority to take, or direct the taking of, all actions and to make all determinations required or provided for under
this Policy and shall have full power and authority to take, or direct the taking of, all such other actions and make all such other determinations
not inconsistent with the specific terms and provisions of this Policy that the Committee deems to be necessary or appropriate to the
administration of this Policy. The interpretation and construction by the Committee of any provision of this Policy and all determinations
made by the Committee under this Policy shall be final, binding and conclusive.
| 9. | Compensation Recovery Repayments not Subject to Indemnification |
Notwithstanding anything to the contrary set forth
in any agreement with, or the organizational documents of, the Company or any of its subsidiaries, Covered Persons are not entitled to
indemnification for Erroneously Awarded Compensation or for any losses arising out of or in any way related to Erroneously Awarded Compensation
recovered under this Policy.
| 10. | No Impairment of Other Remedies |
Nothing contained in this Policy, and no recoupment
or recovery as contemplated herein, shall limit any claims, damages or other legal remedies the Company or any of its affiliates may have
against a Covered Person arising out of or resulting from any actions or omissions by the Covered Person. This Policy does not preclude
the Company from taking any other action to enforce a Covered Person’s obligations to the Company, including, without limitation,
termination of employment and/or institution of civil proceedings. This Policy is in addition to the requirements of Section 304
of the Sarbanes-Oxley Act of 2002 (“SOX 304”) that are applicable to the Company’s Chief Executive Officer and Chief
Financial Officer and to any other compensation recoupment policy and/or similar provisions in any employment, equity plan, equity award,
or other individual agreement, to which the Company is a party or which the Company has adopted or may adopt and maintain from time to
time; provided, however, that compensation recouped pursuant to this Policy shall not be duplicative of compensation recouped pursuant
to SOX 304 or any such compensation recoupment policy and/or similar provisions in any such employment, equity plan, equity award, or
other individual agreement except as may be required by law.
| 11. | Recovery Requirement Shall not Constitute “Good Reason”
Under Employment or Other Compensation Agreements |
Any action by the Company to recoup or any recoupment
of Erroneously Awarded Compensation under this Policy from a Covered Person shall not be deemed (i) “good reason” for
such Covered Person’s resignation or to serve as a basis for a claim of constructive termination under any employment or severance
agreement with the Company or under the terms of any benefits or compensation arrangement applicable to such Covered Person, or (ii) to
constitute a breach of a contract or other arrangement to which such Covered Person is party.
| 12. | Amendment; Termination |
The Committee may amend this Policy in its discretion,
including as it deems necessary to comply with the regulations adopted by the SEC under Rule 10D-1 and the rules of any national
securities exchange or national securities association on which the Company’s securities are listed. The Committee may terminate
this Policy at any time. Notwithstanding anything herein to the contrary, no amendment or termination of this Policy shall be effective
if that amendment or termination would cause the Company to violate any federal securities laws, SEC rules or the rules of any
national securities exchange or national securities association on which the Company’s securities are listed.
This Policy shall be binding and enforceable against
all Covered Persons and their successors, beneficiaries, heirs, executors, administrators, or other legal representatives.
* * *
ACKNOWLEDGMENT
(to be signed by all Covered Persons)
I, the undersigned, agree and acknowledge that I am fully bound by,
and subject to, all of the terms and conditions of the City Therapeutics, Inc. Compensation Recovery Policy (as may be amended, restated,
supplemented or otherwise modified from time to time, the “Policy”) and that I have been provided a copy of the Policy. In
the event of any inconsistency between the Policy and the terms of any employment or similar agreement to which I am a party, or the terms
of any compensation plan, program or agreement under which any compensation has been granted, awarded, earned or paid, the terms of the
Policy shall govern. If the Committee determines that any amounts granted, awarded, earned or paid to me must be forfeited or reimbursed
to the Company, I will promptly take any action necessary to effectuate such forfeiture and/or reimbursement.
Exhibit 10.9
city
THERAPEUTICS, INC.
EXECUTIVE
SEVERANCE PLAN AND
SUMMARY PLAN DESCRIPTION
| 1. | Introduction
and Purpose |
This document serves as the
Plan document and Summary Plan Description (“SPD”) for the severance pay and benefits provided under the City
Therapeutics, Inc. Executive Severance Plan (the “Plan”). City Therapeutics, Inc., a Delaware corporation
(including its successors, the “Company”), considers it essential to foster the continuous employment of key
management personnel. The Company acknowledges the possibility of an involuntary termination of employment exists and that such possibility,
and the uncertainty and questions that it may raise among management, may result in the departure or distraction of management personnel
to the detriment of the Company.
Therefore, the Board has
adopted this Plan to reinforce and encourage the continued attention and dedication of the Covered Executives to their assigned duties
without distraction. This Plan contains information that will help Covered Executives understand the severance pay and benefits being
offered hereunder. The Company encourages all Covered Executives to read the Plan carefully. Note that capitalized words and phrases
used throughout this document are generally defined in Section 4.
Effective as of the date
upon which the registration statement on Form S-1 that is filed by the Company with respect to its initial public offering is declared
effective by the Securities and Exchange Commission (the “Effective Date”), the Board hereby establishes an
unfunded severance benefits plan that is intended to be a welfare benefit plan within the meaning of Section 3(1) of the Employee
Retirement Income Security Act of 1974, as amended (“ERISA”). This Plan, along with the Participant Agreement,
supersedes any and all severance plans, policies and provisions and all change in control plans, policies and provisions applying to
a Covered Executive that may have been in effect before the Effective Date, including by superseding all severance pay, benefits and
equity acceleration provision in any Covered Executive’s offer letter, employment agreement or otherwise.
All Covered Executives who
have executed and submitted to the Company a Participation Agreement, and satisfied such other requirements as may be determined by the
Plan Administrator, are eligible to participate in the Plan. The Plan Administrator may determine
at any time that another employee should be designated as a Covered Executive and shall be eligible to participate in the Plan upon the
Plan Administrator taking action by resolution to update Exhibit A or Exhibit B hereto, as applicable.
For purposes of this Plan,
the following definitions shall apply:
(a) “Accelerated
Vesting Date” means the later of the (i) Covered Executive’s Date of Termination or (ii) effective date
of the Covered Executive’s Separation Agreement and Release (as defined below).
(b) “Accounting
Firm” means a nationally recognized accounting firm selected by the Company.
(c) “Adverse
Benefit Determination” means any of the following: a denial, reduction or termination of, or a failure to provide or make
payment (in whole or in part) for, a benefit, including any such denial, reduction, termination or failure to provide or make payment
that is based on a determination of a Covered Executive’s or beneficiary’s eligibility to participate in the Plan.
(d) “Affiliate”
means any “parent” or “subsidiary” of the Company as such terms are defined in Rule 405 of the Securities
Act of 1933, as amended.
(e) “Base
Salary” means higher of the Covered Executive’s annual base salary in effect immediately prior to (i) the Covered
Executive’s Date of Termination or (ii) a Change in Control.
(f) “Board”
means the Board of Directors of the Plan Sponsor.
(g) “Cause”
means and shall be limited to, the occurrence of any one or more of the following events:
| (i) | the Covered Executive’s material
misconduct with respect to any of the Covered Executive’s duties or responsibilities,
including, without limitation, unlawful harassment or misappropriation of funds or property
of the Company or any of its Affiliates; |
| (ii) | the Covered Executive’s participation
in or commission of any act that would constitute (A) a felony or (B) a crime involving
fraud, dishonesty or moral turpitude under the laws of the United States or any state thereof; |
| (iii) | any conduct by the Covered Executive
that would reasonably be expected to result in material injury to, or material reputational
harm to, the Company or any of its Affiliates if the Covered Executive were retained in the
Covered Executive’s position; |
| (iv) | the Covered Executive’s material
violation of any agreement between the Covered Executive and the Company, any Company policy
or any statutory duty owed to the Company or its stockholders, including, without limitation,
the Continuing Obligations; |
| (v) | an act by the Covered Executive of gross
misconduct or clear and material insubordination; |
| (vi) | the Covered Executive’s commission
or attempted commission of, or participation in, a fraud or act of dishonesty against the
Company; |
| (vii) | the Covered Executive’s continued
failure to perform any of the Covered Executive’s material duties or responsibilities
to the Company (other than by reason of the Covered Executive’s physical or mental
illness, incapacity or disability). |
If the Company terminates the Covered
Executive’s employment other than for Cause, or the Covered Executive resigns, and matters constituting Cause become known to the
Company following the Date of Termination, or if the Covered Executive resigns after the Company learns of matters constituting Cause
but before the Company is able to effect a termination for Cause, the Company may, in either case, by written notice to the Covered Executive,
treat such termination or resignation as a termination for Cause for purposes of this Plan.
Any determination of Cause shall be
made by the Board in its sole discretion, and such determination shall be conclusive for purposes of this Plan
(h) “Change
in Control” means a Sale Event, as defined in the Company’s 2026 Stock Option and Incentive Plan, as amended from
time to time, so long as such Change in Control also constitutes a “change in control event” within the meaning of section
409A of the Code.
(i) “Change
in Control Period” means the period beginning three months prior to a Change in Control and ending on the one-year anniversary
of the Change in Control.
(j) “Code”
means the Internal Revenue Code of 1986, as amended.
(k) “Continuing
Obligations” means the Covered Executive’s obligations to the Company pursuant to any agreement relating to confidentiality,
non-solicitation of customers and employees, assignment of inventions and other restrictive covenants.
(l) “Covered
Executives” means the Tier 1 Executive and those other employees designated by the Plan Administrator in its sole discretion
as the Tier 2 Executives or Tier 3 Executives, in each case, who meet the eligibility requirements set forth in Section 3 of the
Plan.
(m) “Date
of Termination” means the date that a Covered Executive’s employment with the Company and its Affiliates ends, which
date shall be specified in the Notice of Termination. Notwithstanding the foregoing, a Covered Executive’s employment will not
be deemed to have been terminated solely as a result of the Covered Executive becoming an employee of any direct or indirect successor
to the business or assets of the Company or any Affiliate.
(n) “Disability”
means the Covered Executive is disabled and unable to perform or expected to be unable to perform the essential functions of the Covered
Executive’s then existing position or positions with or without reasonable accommodation for a period of 180 days (which need not
be consecutive) in any 12-month period.
(o) “Good
Reason” means the Covered Executive’s resignation from employment with the Company within twelve (12) months after
any of the following conditions has come into existence without the Covered Executive’s consent:
| (i) | a reduction of the Covered Executive’s
base salary by ten percent (10%) or more without the Covered Executive’s consent; |
| (ii) | a material diminution of the Covered
Executive’s authority, duties or responsibilities; or |
| (iii) | a breach by the Company of the terms
of the Covered Executive’s offer letter or other written employment agreement with
the Company. |
For purposes of this definition, Good
Reason shall not be deemed to have occurred unless the Covered Executive gives the Company written notice of the applicable condition
within ninety (90) days after such condition comes into existence, the Company fails to remedy the condition within thirty (30) days
after receiving such written notice (the “Cure Period”), and the Covered Executive terminates the Covered Executive’s
employment within sixty (60) days after the end of the Cure Period.
(p) “Notice”
or “Notification” means the delivery or furnishing of information to an individual in a manner that is reasonably
calculated to ensure actual receipt by the individual.
(q) “Notice
of Termination” means a written Notice that indicates the specific termination provision in this Plan relied upon for the
termination of a Covered Executive’s employment and the Date of Termination.
(r) “Participation
Agreement” shall mean an agreement between a Covered Executive and the Company that acknowledges
and agrees to the Covered Executive’s participation in the Plan and to any related terms set forth in therein.
(s) “Plan
Administrator” means the Board or a committee thereof designated by the Board to administer the Plan; provided,
however, that the Plan Administrator may in its sole discretion appoint a new Plan Administrator to administer the Plan at any
time.
(t) “Plan
Sponsor” means the Company.
(u) “Qualified
Termination” means (i) a termination of the Covered Executive’s employment by the Company other than for Cause,
death or Disability or (ii) the Covered Executive’s resignation from the Company for Good Reason.
(v) “Separation
Obligations” means: (i) execution of a separation agreement and release in a form and manner satisfactory to and provided
by the Company that contains, among other provisions, a confirmation of the Covered Executive’s resignation from all officer and
director positions with the Company and any Affiliate as of the Date of Termination, a general release of claims in favor of the Company
and related persons and entities, confidentiality, return of property and non-disparagement provisions, a reaffirmation of the Covered
Executive’s Continuing Obligations and provides that if the Covered Executive breaches any of the Continuing Obligations, all severance
payments and benefits shall immediately cease (the “Separation Agreement and Release”) and (ii) the Separation
Agreement and Release becoming irrevocable, all within 60 days after the Date of Termination (or such shorter period set forth in the
Separation Agreement and Release).
(w) “Target
Bonus” means the higher of the Covered Executive’s target annual cash incentive compensation or target sales incentive
plan commission, as applicable, in effect immediately prior to (i) the Covered Executive’s Date of Termination or (ii) the
Change in Control.
(x) “Tier
1 Executive” means the Company’s Chief Executive Officer.
(y) “Tier
2 Executives” means the individuals holding the titles designated as such by the Plan Administrator and whose titles are
listed in Exhibit A, attached hereto, as such exhibit may be amended by the Plan Administrator from time to time.
(z) “Tier
3 Executives” means the individuals holding the titles designated as such by the Plan Administrator and whose titles are
listed in Exhibit B, attached hereto, as such exhibit may be amended by the Plan Administrator from time to time.
(aa) “Time-Based
Equity Awards” means all outstanding stock options, restricted stock units, restricted stock awards and other stock-based
awards with respect to the Company’s common stock that are subject solely to time-based vesting.
If the Covered Executive’s
employment with the Company is terminated for any reason, the Covered Executive shall be deemed to have resigned from any officer or
director positions that the Covered Executive holds with the Company or any Affiliate as of the Date of Termination. the Company shall
pay or provide to the Covered Executive (or to the Covered Executive’s authorized representative or estate) (i) any base salary
earned but unpaid through the Date of Termination; (ii) unpaid expense reimbursements (subject to, and in accordance with, the Company’s
expense reimbursement policy then in effect); and (iii) any vested benefits the Covered Executive may have under any employee benefit
plan of the Company through the Date of Termination, which vested benefits shall be paid and/or provided in accordance with the terms
of such employee benefit plans (collectively, the “Accrued Obligations”).
| 6. | Qualified
Termination not in Connection with a Change in Control |
If a Covered Executive experiences
a Qualified Termination outside of the Change in Control Period, then, in addition to the Accrued Obligations, and subject to the Covered
Executive’s fulfillment of the Separation Obligations:
(a) the
Company shall pay to the Covered Executive an amount equal to 12 months’ Base Salary for the Tier 1 Executive, 12 months’
Base Salary for a Tier 2 Executive and 9 months’ Base Salary for a Tier 3 Executive; and
(b) subject
to the Covered Executive’s proper election to receive COBRA benefits, the Company shall pay to the group health plan provider or
the COBRA provider a monthly payment equal to the employer portion of the monthly COBRA premium until the earliest of (i) the 12-month
anniversary for the Tier 1 Executive, the 12-month anniversary for a Tier 2 Executive and the nine-month anniversary for a Tier 3 Executive,
in each case, following the Date of Termination; (ii) the date that the Covered Executive becomes eligible for group medical plan
benefits under another employer’s group medical plan; or (iii) the cessation of the Covered Executive’s health continuation
rights under COBRA; provided, however, that if the Company determines that such payments may violate applicable law, the
Company shall instead make such payments directly to the Covered Executive in the form of payroll.
The amounts payable under
Section 6(a) and 6(b), as applicable, will be paid out in substantially equal installments in accordance with the Company’s
payroll practice over 12 months in the case of the Tier I Executive or a Tier 2 Executive, or nine months in the case of a Tier 3 Executive,
commencing within 60 days after the Date of Termination; provided, however, that if the 60-day period begins in one calendar
year and ends in a second calendar year, such payments, to the extent they qualify as “non-qualified deferred compensation”
within the meaning of Section 409A of the Code, shall begin to be paid in the second calendar year by the last day of such 60-day
period; provided, further, that the initial payment shall include a catch-up payment to cover amounts retroactive to the
day immediately following the Date of Termination.
If any Covered Executive
experiences a Qualified Termination within the Change in Control Period, then, in addition to the Accrued Obligations, and subject to
the Covered Executive’s fulfillment of the Separation Obligations:
(a) the
Company shall pay to the Covered Executive an amount equal to the sum of (i) 150% of Base Salary for the Tier 1 Executive, 100%
of Base Salary for a Tier 2 Executive and 75% of Base Salary for a Tier 3 Executive and (ii) 150% of the Target Bonus for the Tier
1 Executive, 100% of the Target Bonus for a Tier 2 Executive and 75% of the Target Bonus for a Tier 3 Executive;
(b) subject
to the Covered Executive’s proper election to receive COBRA benefits, the Company shall pay to the group health plan provider or
the COBRA provider a monthly payment equal to the monthly COBRA premium until the earliest of (i) the 18-month anniversary for the
Tier 1 Executive, the 12-month anniversary for a Tier 2 Executive and the nine-month anniversary for a Tier 3 Executive, in each case,
following the Date of Termination; (ii) the date that the Covered Executive becomes eligible for group medical plan benefits under
another employer’s group medical plan; or (iii) the cessation of the Covered Executive’s health continuation rights
under COBRA; provided, however, that if the Company determines that such payments may violate applicable law, the Company
shall instead make such payments directly to the Covered Executive in the form of payroll; and
(c) notwithstanding
anything to the contrary in any applicable Time-Based Equity Award agreement or in any applicable Company equity incentive plan, all
Time-Based Equity Awards shall immediately accelerate and become fully vested and exercisable or nonforfeitable as of the Accelerated
Vesting Date, provided that in order to effectuate the accelerated vesting contemplated by this subsection, the unvested portion of the
Covered Executive’s Time-Based Equity Awards that would otherwise terminate or be forfeited on the Date of Termination will be
delayed until the earlier of (i) the effective date of the Separation Agreement and Release or the date of the Change in Control,
as applicable (at which time acceleration will occur), or (ii) the date that the Separation Agreement and Release can no longer
become fully effective (at which time the unvested portion of the Covered Executive’s Time-Based Equity Awards will terminate or
be forfeited). Notwithstanding the foregoing, no additional vesting of the Time-Based Equity Awards will occur during the period between
the Date of Termination and the Accelerated Vesting Date. For the avoidance of doubt, upon a Qualifying Termination prior to the consummation
of a Change in Control (1) all then unvested Time-Based Equity Awards held by the Covered Executive as of the Date of Termination
will remain outstanding for a period of three months thereafter (unless such Time-Based Equity Awards are forfeited prior to such date
pursuant to clause (ii) of the preceding sentence) and remain eligible to become vested upon the consummation of such Change in
Control and, to the extent such Time-Based Equity Awards do not become vested in accordance with the terms set forth in this Section 7(c) on
or prior to the date that is three months following the Date of Termination, all then-unvested Time-Based Equity Awards held by such
Covered Executive will automatically and without further action be canceled and forfeited on the three-month anniversary of the Date
of Termination.
The amounts payable under
Section 7(a) shall be paid out in a lump sum within 60 days after the Date of Termination and the amounts payable under Section 7(b) will
be paid out in substantially equal installments in accordance with the Company’s payroll practice over 18 months in the case of
the Tier 1 Executive, 12 months in the case of a Tier 2 Executive or nine months in the case of a Tier 3 Executive, commencing within
60 days after the Date of Termination; provided, however, that if the 60-day period begins in one calendar year and ends
in a second calendar year, such payments, to the extent they qualify as “non-qualified deferred compensation” within the
meaning of Section 409A of the Code, shall begin to be paid in the second calendar year by the last day of such 60-day period; provided,
further, that the initial payment shall include a catch-up payment to cover amounts retroactive to the day immediately following
the Date of Termination.
For the avoidance of doubt,
(i) in no event will the Covered Executive be entitled to severance payments and benefits under both Section 6 and Section 7
of this Plan and (ii) if a Covered Executive’s Date of Termination occurs during the Change in Control Period but prior to
a Change in Control (x) any severance pay and benefits previously provided to the Covered Executive under Section 6 shall reduce
the severance pay and benefits to be provided under this Section 7 and (y) payment of any overlapping amounts under Section 7(a)(i) above
shall continue to be paid in installments pursuant to the schedule specified in Section 6 to the extent necessary to comply with
Section 409A of the Code.
(a) This
Plan is intended to be administered in accordance with Section 409A of the Code. To the extent that any provision of this Plan is
ambiguous as to its compliance with Section 409A of the Code, the provision shall be read in such a manner so that all payments
hereunder comply with Section 409A of the Code. Each payment pursuant to this Plan is intended to constitute a separate payment
for purposes of Treasury Regulation Section 1.409A-2(b)(2). This Plan may be amended as may be necessary to fully comply with Section 409A
of the Code and all related rules and regulations in order to preserve the payments and benefits provided hereunder without additional
cost.
(b) To
the extent that any payment or benefit described in this Plan constitutes “non-qualified deferred compensation” under Section 409A
of the Code, and to the extent that such payment or benefit is payable upon the Covered Executive’s termination of employment,
then such payments or benefits shall be payable only upon the Covered Executive’s “separation from service.” The determination
of whether and when a separation from service has occurred shall be made in accordance with the presumptions set forth in Treasury Regulation
Section 1.409A-1(h).
(c) Anything
in this Plan to the contrary notwithstanding, if at the time of the Covered Executive’s separation from service within the meaning
of Section 409A of the Code, the Company determines that the Covered Executive is a “specified employee” within the
meaning of Section 409A(a)(2)(B)(i) of the Code, then to the extent any payment or benefit that the Covered Executive becomes
entitled to under this Plan on account of the Covered Executive’s separation from service would be considered deferred compensation
otherwise subject to the 20% additional tax imposed pursuant to Section 409A(a) of the Code as a result of the application
of Section 409A(a)(2)(B)(i) of the Code, such payment shall not be payable and such benefit shall not be provided until the
date that is the earlier of (A) six months and one day after the Covered Executive’s separation from service, or (B) the
Covered Executive’s death. If any such delayed cash payment is otherwise payable on an installment basis, the first payment shall
include a catch-up payment covering amounts that would otherwise have been paid during the six-month period but for the application of
this provision (without interest), and the balance of the installments shall be payable in accordance with their original schedule.
(d) The
Company makes no representation or warranty and shall have no liability to any Covered Executive or any other person if any provisions
of this Plan are determined to constitute deferred compensation subject to Section 409A of the Code but do not satisfy an exemption
from, or the conditions of, such Section.
(a) Anything
in this Plan to the contrary notwithstanding, in the event that the amount of any compensation, payment or distribution by the Company
to or for the benefit of the Covered Executive, whether paid or payable or distributed or distributable pursuant to the terms of this
Plan or otherwise, calculated in a manner consistent with Section 280G of the Code and the applicable regulations thereunder (the
“Aggregate Payments”), would be subject to the excise tax imposed by Section 4999 of the Code, then the
Aggregate Payments shall be reduced (but not below zero) so that the sum of all of the Aggregate Payments shall be $1.00 less than the
amount at which the Covered Executive becomes subject to the excise tax imposed by Section 4999 of the Code; provided that such
reduction will only occur if it would result in the Covered Executive receiving a higher After Tax Amount (as defined below) than the
Covered Executive would receive if the Aggregate Payments were not subject to such reduction. Notwithstanding the foregoing, if, immediately
before the change in ownership or control, no stock of the Company is readily tradeable on an established securities market or otherwise
(determined in accordance with Q&A 6 of Treasury Regulations Section 1.280G-1), the Company shall use reasonable efforts to
satisfy the shareholder approval requirements set forth in Q&A 7 of Treasury Regulations Section 1.280G-1 with respect to the
amount of any such potential reduction, and if such requirements are satisfied then no such reduction will apply. In the event of a reduction,
the Aggregate Payments will be reduced in the following order, in each case, in reverse chronological order beginning with the Aggregate
Payments that are to be paid the furthest in time from consummation of the transaction that is subject to Section 280G of the Code:
(i) cash payments not subject to Section 409A of the Code; (ii) cash payments subject to Section 409A of the Code;
(iii) equity-based payments and acceleration; and (iv) non-cash forms of benefits; provided that in the case of all the foregoing
Aggregate Payments all amounts or payments that are not subject to calculation under Treas. Reg. §1.280G-1, Q&A-24(b) or
(c) shall be reduced before any amounts that are subject to calculation under Treas. Reg. §1.280G-1, Q&A-24(b) or
(c).
(b) For
purposes of this Section, the “After Tax Amount” means the amount of the Aggregate Payments less all federal,
state and local income, excise, employment and social security taxes imposed on the Covered Executive as a result of the Covered Executive’s
receipt of the Aggregate Payments. For purposes of determining the After Tax Amount, the Covered Executive shall be deemed to pay federal
income taxes at the highest marginal rate of federal income taxation applicable to individuals for the calendar year in which the determination
is to be made, and state and local income taxes and social security taxes at the highest marginal rates of individual taxation in each
applicable state and locality, net of the maximum reduction in federal income taxes (if any) that could be obtained from deduction of
such state and local taxes.
(c) The
determination as to whether a reduction in the Aggregate Payments shall be made pursuant to Section 9(a) shall be made by the
Accounting Firm selected by the Company prior to the closing of the Change in Control, which shall provide detailed supporting calculations
both to the Company and the Covered Executive within 15 business days of the Date of Termination, if applicable, or at such earlier time
as is reasonably requested by the Company or the Covered Executive. Any determination by the Accounting Firm shall be binding upon the
Company and the Covered Executive.
| 10. | Withholding;
Tax Effect |
All payments made under this
Plan shall be net of any tax or other amounts required to be withheld by the Company under applicable law. Nothing in this Plan shall
be construed to require the Company to make any payments to compensate the Covered Executive for any adverse tax effect associated with
any payments or benefits or for any deduction or withholding from any payment or benefit.
If a Covered Executive dies
after the Date of Termination, but before all payments or benefits to which such Covered Executive is entitled pursuant to the Plan have
been paid or provided, any remaining payments and benefits will be made to any beneficiary designated by the Covered Executive prior
to the Covered Executive’s death (or the Covered Executive’s estate, if the Covered Executive fails to make such designation).
(a) Plan
Administrator. The Plan Administrator and Plan Sponsor shall serve as the “named fiduciaries” of the Plan under ERISA.
The Plan Administrator shall be the “administrator” within the meaning of Section 3(16) of ERISA and shall have all
the responsibilities and duties contained therein.
(b) Decisions,
Powers and Duties. The general administration of the Plan and the responsibility for carrying out its provisions shall be vested
in the Plan Administrator. The Plan Administrator shall have the maximum discretionary authority permitted by law to discharge such duties
and responsibilities, which also include, but are not limited to, interpretation and construction of the Plan, the determination of all
questions of fact, including, without limitation, eligibility, participation and benefits, the resolution of any ambiguities and all
other related or incidental matters, and such duties and powers of the Plan administration that are not assumed from time to time by
any other appropriate entity, individual or institution. The Plan Administrator may adopt rules and regulations of uniform applicability
in its interpretation and implementation of the Plan.
The Plan Administrator shall
discharge its duties and responsibilities and exercise its powers and authority in its sole discretion and in accordance with the terms
of the controlling legal documents and applicable law, and its actions and decisions that are not arbitrary and capricious shall be binding
on any Covered Executive, and the Covered Executive’s spouse or other dependent or beneficiary and any other interested parties
whether or not in being or under a disability.
(c) Reports.
The Plan Administrator shall be responsible for the preparation and delivery of all reports, notices, plan summaries and plan descriptions
required to be filed with any governmental office or to be given to any employee, former employee or beneficiary.
(d) Company
to Furnish Information. Upon request of the Plan Administrator, the Company shall furnish such information in its possession and
aid the Plan Administrator in the performance of its duties hereunder.
| 13. | Claims
Procedure and Payment of Benefits |
(a) Application
for Benefits
(i) General.
All applications for benefits under the Plan shall be submitted to the Plan Administrator at such location as designated by the Plan
Administrator from time to time. Applications for benefits must be in writing on forms acceptable to the Plan Administrator and must
be signed by the Covered Executive. The Plan Administrator reserves the right to require the Covered Executive to furnish such other
information and documents as the Plan Administrator determines are necessary or appropriate. Each application shall be acted upon and
approved or disapproved by the Plan Administrator within 90 days following its receipt by the Plan Administrator.
(ii) Notification.
The Plan Administrator shall provide a claimant with written or electronic Notification of any Adverse Benefit Determination. Any electronic
Notification shall comply with the standards imposed by 29 C.F.R. 2520.104b-l(c)(l)(i), (iii) and (iv). The Notification shall set
forth, in a manner calculated to be understood by the claimant:
| (A) | the
specific reason or reasons for the Adverse Benefit Determination; |
| (B) | reference
to the specific Plan provisions on which the Adverse Benefit Determination is based; |
| (C) | a
description of any additional material or information necessary for the claimant to perfect
the claim and an explanation of why such material or information is necessary; and |
| (D) | a
description of the Plan’s review procedures and the time limits applicable to such
procedures, including a statement of the claimant’s right to bring a civil action under
Section 502(a) of ERISA following an Adverse Benefit Determination on review. |
(b) Review
of Benefit Denials
(i) Appeal.
A claimant shall have a reasonable opportunity to appeal an Adverse Benefit Determination to the Board and under which there will be
a full and fair review of the claim and the Adverse Benefit Determination. The Board shall be identified in the Notification described
in Section 13(a)(ii) and may be the Plan Administrator. Such full and fair review shall:
| (A) | provide
claimants at least 60 days following receipt of a Notification of an Adverse Benefit Determination
within which to appeal the Adverse Benefit Determination; |
| (B) | provide
claimants the opportunity to submit written comments, documents, records and other information
relating to the claim for benefits; |
| (C) | provide
that a claimant shall be provided, upon request and free of charge, reasonable access to,
and copies of, all documents, records and other information relevant to the claimant’s
claim for benefits; and |
| (D) | provide
for a review of the initial Adverse Benefit Determination that takes into account all comments,
documents, records and other information submitted by the claimant relating to the claim,
without regard to whether such information was submitted or considered in the initial Adverse
Benefit Determination. |
(ii) Timing
of Benefit Determination on Review. The Board shall notify the Covered Executive, in accordance with Section 13(b)(iv), of the
Plan Administrator’s benefit determination on review within a reasonable period of time. Such Notification shall be provided not
later than 60 days after receipt by the Plan Administrator of the Covered Executive’s request for review of the Adverse Benefit
Determination.
(iii) Furnishing
Documents. In the case of an Adverse Benefit Determination on review, the Board shall provide such access to, and copies of, documents,
records and other information described in Section 13(b)(iv) as is appropriate.
(iv) Content
of Notification on Review. The Board shall provide a claimant with written or electronic Notification of a Plan Administrator’s
benefit determination following an appeal described in Section 13(b)(i). Any electronic Notification shall comply with the standards
imposed by 29 C.F.R. 2520.104b-l(c)(l)(i), (iii) and (iv). In the case of an Adverse Benefit Determination on review, the Notification
shall set forth, in a manner calculated to be understood by the claimant:
| (A) | the
specific reason or reasons for the Adverse Benefit Determination; |
| (B) | reference
to the specific Plan provisions on which the Adverse Benefit Determination is based; |
| (C) | a
statement that the claimant is entitled to receive, upon request and free of charge, reasonable
access to, and copies of, all documents, records and other information relevant to the claimant’s
claim for benefits; and |
| (D) | a
statement of the claimant’s right to bring an action under Section 502(a) of
ERISA. |
(c) Calculating
Time Periods. The period of time within which a benefit determination (including an Adverse Benefit Determination on review) is required
to be made shall begin at the time a claim (or appeal) is filed in accordance with the reasonable procedures of the Plan, without regard
to whether all the information necessary to make a benefit determination accompanies the filing. In the event that a period of time is
extended due to a claimant's failure to submit information necessary to decide a claim, the period for making the benefit determination
shall be tolled from the date on which the Notification of the extension is sent to the claimant until the date on which the claimant
responds to the request for additional information.
(d) Extension
of Notice Period. The 90 day and 60 day periods applicable to the Notice furnished by the Plan Administrator in Sections 13(a) and
13(b) above may be extended at the discretion of the Plan Administrator for a second 90 or 60 day period, as the case may be, provided
that written Notice of the extension is furnished to the claimant prior to the termination of the initial period, indicating the special
circumstances requiring such extension of time and the date by which a final decision is expected.
(e) Facility
of Payment. Whenever and as often as any person entitled to payments hereunder shall be determined to be a minor or under other legal
disability or otherwise incapacitated in any way so as to be unable to manage the person’s financial affairs, the Company, in its
discretion, may direct that all or any portion of the benefit payments be made: (i) to such person; (ii) to such person’s
legal guardian or conservator; or (iii) to such person’s spouse. The decision of the Plan Administrator shall, in each case,
be final and binding upon all persons. Any payment made pursuant to the authority herein conferred shall operate as a complete discharge
of the obligations of the Company under the Plan in respect thereof.
(f) Responsibility
for Payment. The Company shall be liable for the payment of benefits in accordance with the terms of the Plan. The benefits under
the Plan shall be payable solely by the Company and each Covered Executive who shall claim the right to any payment under the Plan shall
be entitled to look only to the Company for such payment.
(g) Limitation
on Benefits. The Company shall have no obligation to set aside, earmark or entrust any fund, policy or money with which to pay its
obligations under this Plan. The Covered Executive, or any successor in interest, shall be and remain simply a general, unsecured creditor
of the Company with respect to the benefits under this Plan in the same manner as any other creditor who has a general claim for an unpaid
liability.
To the extent permitted by
law, all employees, officers, directors, agents and representatives of the Company shall be indemnified by the Company and held harmless
against any claims and the expenses of defending against such claims, resulting from any action or conduct relating to the administration
of the Plan, whether as a member of the Plan Administrator or otherwise, except to the extent that such claims arise from gross negligence,
willful neglect or willful misconduct.
| 15. | Plan;
At Will Employment |
The Plan is not a contract
between the Company and any employee, nor is it a condition of employment or services of any employee. Nothing contained in the Plan
gives, or is intended to give, any employee the right to be retained in the service of the Company, or to interfere with the right of
the Company to discharge or terminate the employment or services of any employee at any time and for any reason. No employee shall have
the right or claim to benefits beyond those expressly provided in this Plan, if any. All rights and claims are limited as set forth in
the Plan.
If any portion or provision
of this Plan (including, without limitation, any portion or provision of any Section of this Plan) is to any extent declared illegal
or unenforceable by a court of competent jurisdiction, then the remainder of this Plan, or the application of such portion or provision
in circumstances other than those as to which it is so declared illegal or unenforceable, shall not be affected thereby, and each portion
and provision of this Plan shall be valid and enforceable to the fullest extent permitted by law.
No waiver of any provision
hereof shall be effective unless made in writing and signed by the waiving party. The failure of any party to require the performance
of any term or obligation of this Plan, or the waiver by any party of any breach of this Plan, shall not prevent any subsequent enforcement
of such term or obligation or be deemed a waiver of any subsequent breach.
| 18. | Successors
and Assigns |
No right or interest of any
Covered Executive in the Plan shall be assignable or transferable in whole or in part either directly or by operation of law or otherwise,
including, but not limited to, execution, levy, garnishment, attachment, pledge or bankruptcy. The Plan Sponsor may assign or otherwise
transfer this Plan without any Covered Executive’s consent to any affiliate or to any person or entity with whom the Plan Sponsor
shall hereafter effect a reorganization or consolidation, into which the Plan Sponsor merges or to whom it transfers all or substantially
all of its properties or assets; provided that if the Covered Executive remains employed or becomes employed by the Company, the
purchaser or any of their affiliates in connection with any such transaction, then the Covered Executive shall not be entitled to any
payments or benefits pursuant to Sections 6 or 7 of this Plan solely as a result of such transaction. This Plan shall inure to the benefit
of and be binding upon the Covered Executive and the Plan Sponsor, and each of the Covered Executive’s and the Plan Sponsor’s
respective successors, executors, administrators, heirs and permitted assigns.
| 19. | Non-Duplication
of Benefits and Effect on Other Plans |
Notwithstanding any other
provision in the Plan to the contrary, the benefits provided hereunder and, if applicable, pursuant to the Participant Agreement, are
in lieu of any other severance payments and/or benefits provided by the Company, including any such payments and/or benefits pursuant
to an employment agreement or offer letter between the Company and any Covered Executive.
| 20. | Amendment
or Termination |
The Plan Sponsor may amend
or terminate this Plan at any time or from time to time, but no such action shall adversely affect the rights of any Covered Executive
who has signed a Participation Agreement without such Covered Executive’s written consent.
| 21. | Governing
Law and Limitation on Actions |
The Plan and the rights of
all persons hereunder shall be construed in accordance with and under applicable provisions of ERISA, and the regulations thereunder,
and the Plan and the rights of all persons under the Plan shall be construed in accordance with and under applicable provisions the laws
of the Commonwealth of Massachusetts (without regard to conflict of laws provisions), to the extent not preempted by federal law. No
action (whether at law, in equity or otherwise) shall be brought by or on behalf of any person for or with respect to benefits due under
this Plan unless the person bringing such action has timely exhausted the Plan’s claim review procedure. Any action (whether at
law, in equity or otherwise) must be filed and litigated in the U.S. District Court for the District of Massachusetts, and must be commenced
within one year. This one-year period shall be computed from the earlier of (a) the date a final determination denying such benefit,
in whole or in part, is issued under the Plan’s claim review procedure and (b) the date such person’s cause of action
first accrued. If a person does not bring such an action within such one-year period, the person will be barred from bringing such action.
| 22. | ERISA
Required Information |
Official Plan Name: The official name
of the Plan is the City Therapeutics, Inc. Executive Severance Plan.
Plan Administrator: The Plan Administrator
is the Board (or its designee) of the Plan Sponsor.
Plan Sponsor:
City Therapeutics, Inc.
399 Binney Street
Cambridge, MA 02142
(857) 357-2024
Plan Number: Documents and reports for
the Plan are filed with the U.S. Department of Labor under City Therapeutics, Inc. Employer Identification Number (EIN) and the
Plan Number (PN). The EIN for City Therapeutics, Inc. is 93-2762122, and the PN is 502.
The Plan Year: The Plan year is January 1
through December 31.
Agent for Service of Legal Process: City
Therapeutics, Inc. is the Plan’s agent for service of legal process and may be served at the address indicated in the section
entitled “Plan Administrator and Sponsor” above.
Funding: All severance pay and benefits
are paid from the Company’s general assets and, thus the Plan is considered unfunded for tax purposes under Title I of ERISA.
ERISA Rights: The Plan is subject to ERISA.
As a Covered Executive, you are entitled to certain rights and protections under ERISA. Under ERISA, you are entitled to:
| · | Examine,
without charge at the Plan Administrator’s office, all official Plan documents (including
insurance contracts) and copies of all documents filed with the U.S. Department of Labor,
such as detailed Summary Annual Reports. |
| · | Obtain
copies of all official Plan documents and other Plan information upon written request to
the Plan Administrator. The Plan Administrator may charge a reasonable fee for the copies. |
| · | Receive
a summary of the Plan’s annual financial report. The Plan Administrator is required
to furnish each participant with a copy of this Summary Annual Report. |
You also have the right to expect “fiduciaries,”
the people who are responsible for the management of the Plan, to act prudently and to act in the interest of you and other Plan participants
and beneficiaries. Another one of your ERISA-guaranteed rights means that no one – including the Company or any other person –
may fire you or otherwise discriminate against you in any way to prevent you from obtaining a Plan benefit or exercising your rights
under ERISA.
ERISA also guarantees your rights to written
notice if any part of a claim is denied or ignored, in whole or in part. Because your rights under ERISA are protected by law, you also
can file suit if a right is denied. For example, if you request certain Plan-related materials from the Plan Administrator and do not
receive them within 30 days, you may file suit in a federal court. In such a case, the court may require the Plan Administrator to provide
the materials and pay a fine of up to $110 a day until you receive the materials, unless the materials were not sent because of reasons
beyond the control of the Plan Administrator. You may file suit in a federal court if you have a claim for benefits under the Plan that
is denied or ignored, in whole or in part. You also can seek assistance from the U.S. Department of Labor or file suit in a federal court
if a Plan fiduciary has misused Plan funds or if you are discriminated against for asserting your rights. The court will decide who should
pay court costs and legal fees. If you are successful, the court may order the person you have sued to pay these costs and fees. If you
lose because, for example, the court finds your claim frivolous, you may be ordered to pay all these costs and fees on your own, including
any court costs and attorney fees.
If you have any questions about the Plan, you
should contact the Plan Administrator. If you have any questions about this statement or about your rights under ERISA, or if you need
assistance in obtaining documents from the Plan Administrator, you should contact the nearest office of the Employee Benefits Security
Administration, U.S. Department of Labor, listed in your telephone directory or the Division of Technical Assistance and Inquiries, Employee
Benefits Security Administration, U.S. Department of Labor, 200 Constitution Ave. N.W., Washington, D.C. 20210. You may also obtain certain
publications about your rights and responsibilities under ERISA by calling the Employee Benefits Security Administration’s publications
hotline at 1-866-444-3272.
Exhibit A
Tier 2 Executives
| Title |
| Chief
Financial Officer |
| Chief
Medical Officer |
| Chief
Scientific Officer |
| Chief
Operating Officer |
| Chief
Legal Officer |
| Chief
Human Resources Officer |
Exhibit B
Tier 3 Executives
| Title |
| All
Company Employees at the Senior Vice President level |
[DATE]
[NAME]
[ADDRESS]
[ADDRESS]
Re: Executive Severance Plan
Dear [NAME],
City Therapeutics, Inc., a Delaware corporation
(the “Company”) is pleased to inform you that you have been designated as an eligible participant in the Company’s
Executive Severance Plan, as amended from time to time (the “Severance Plan”), a copy of which (excluding the exhibits thereto)
is attached hereto as Exhibit A. You have been designated as a Tier [2][3] Executive under the Severance Plan.
Under certain circumstances, you will be eligible
for certain severance benefits as described in the Severance Plan. Any and all such severance benefits are subject to the terms and conditions
of the Severance Plan.
As a condition to participate in the Severance
Plan, you hereby acknowledge that the severance benefits that may be provided to you under the Severance Plan will supersede and replace
any severance benefit plan, policy or practice previously maintained by the Company or any of its affiliates that may have been applicable
to you and any severance benefits under any individually negotiated employment agreement, offer letter agreement or equity award agreement
between you and the Company or any of its affiliates, as may be amended from time to time. In addition, as a condition to participate
in the Severance Plan, you hereby acknowledge that you will continue to comply with the Proprietary Information and Inventions Agreement
entered into between you and the Company on [DATE].
Please review the information in this letter
and the Severance Plan carefully. If you have any questions regarding the letter or the Severance Plan, please contact Anna O’Driscoll
at aodriscoll@citytx.com.
To accept the terms of this letter and participate
in the Severance Plan, please sign and date this letter in the space provided below and return the signed copy to Anna O’Driscoll
by [DATE] (the “Expiration Date”). If you do not return the signed copy by the Expiration Date, the terms of this letter
shall be null and void and you may not participate in the Severance Plan.
| City Therapeutics, Inc. |
|
| |
|
| |
|
| Name: |
|
| Title: |
|
| |
|
| Agreed and Accepted: |
|
| |
|
| |
|
| Name: |
|
| Date: |
|
Exhibit A
City Therapeutics, Inc. Executive Severance
Plan
Exhibit 23.1
CONSENT OF INDEPENDENT REGISTERED PUBLIC ACCOUNTING FIRM
We hereby consent to the use in this Registration Statement on Form
S-1 of City Therapeutics, Inc. of our report dated July 14, 2026, except for the effects of the stock split discussed in Note 18 to the
consolidated financial statements, as to which the date is October 9, 2026, relating to the financial statements of City Therapeutics,
Inc., which appears in this Registration Statement. We also consent to the reference to us under the heading “Experts” in
such Registration Statement.
/s/ PricewaterhouseCoopers LLP
Boston, Massachusetts
October 9, 2026
EX-FILING FEES
S-1
S-1/A
EX-FILING FEES
333-299117
0002143850
CITY THERAPEUTICS, INC.
N/A
N/A
0002143850
2026-10-08
2026-10-08
0002143850
1
2026-10-08
2026-10-08
0002143850
2
2026-10-08
2026-10-08
iso4217:USD
xbrli:pure
xbrli:shares
|
Calculation of Filing Fee Tables
|
|
S-1
|
|
CITY THERAPEUTICS, INC.
|
|
Table 1: Newly Registered and Carry Forward Securities
|
☐Not Applicable
|
|
|
|
Security Type
|
Security Class Title
|
Fee Calculation or Carry Forward Rule
|
Amount Registered
|
Proposed Maximum Offering Price Per Unit
|
Maximum Aggregate Offering Price
|
Fee Rate
|
Amount of Registration Fee
|
Carry Forward Form Type
|
Carry Forward File Number
|
Carry Forward Initial Effective Date
|
Filing Fee Previously Paid in Connection with Unsold Securities to be Carried Forward
|
|
Newly Registered Securities
|
|
Fees to be Paid
|
1
|
Equity
|
Common Stock, $0.0001 par value per share
|
457(a)
|
5,917,397
|
$
19.00
|
$
112,430,543.00
|
0.000087
|
$
9,781.46
|
|
|
|
|
|
Fees Previously Paid
|
2
|
Equity
|
Common Stock, $0.0001 par value per share
|
457(o)
|
5,263,158
|
$
19.00
|
$
100,000,000.00
|
|
$
13,810.00
|
|
|
|
|
|
Carry Forward Securities
|
|
Carry Forward Securities
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
|
Total Offering Amounts:
|
|
$
212,430,543.00
|
|
$
23,591.46
|
|
|
|
|
|
|
|
|
Total Fees Previously Paid:
|
|
|
|
$
13,810.00
|
|
|
|
|
|
|
|
|
Total Fee Offsets:
|
|
|
|
$
0.00
|
|
|
|
|
|
|
|
|
Net Fee Due:
|
|
|
|
$
9,781.46
|
|
|
|
|
|
1
|
(a) Maximum Aggregate Offering Price estimated solely for the purpose of calculating the registration fee pursuant to Rule 457(a) under the Securities Act of 1933, as amended. (b) Includes the aggregate offering price of 1,458,333 additional shares that the underwriters have the option to purchase from the Registrant, if any.
|
|
|
2
|
(a) The Registrant previously paid a registration fee of $13,810.00 in connection with initial filing of the Registration Statement on Form S-1 on September 24, 2026. The fee was estimated solely for the purpose of calculating the registration fee in accordance with Rule 457(o) under the Securities Act of 1933, as amended. This Maximum Aggregate Offering Price was originally registered under 457(o) and is now converted to 457(a). (b) See note 1(b) above.
|
|
|
Table 2: Fee Offset Claims and Sources
|
☑Not Applicable
|
|
|
|
Registrant or Filer Name
|
Form or Filing Type
|
File Number
|
Initial Filing Date
|
Filing Date
|
Fee Offset Claimed
|
Security Type Associated with Fee Offset Claimed
|
Security Title Associated with Fee Offset Claimed
|
Unsold Securities Associated with Fee Offset Claimed
|
Unsold Aggregate Offering Amount Associated with Fee Offset Claimed
|
Fee Paid with Fee Offset Source
|
|
Rules 457(b) and 0-11(a)(2)
|
|
Fee Offset Claims
|
|
|
|
|
|
|
|
|
|
|
|
|
|
Fee Offset Sources
|
|
|
|
|
|
|
|
|
|
|
|
|
|
Rule 457(p)
|
|
Fee Offset Claims
|
|
|
|
|
|
|
|
|
|
|
|
|
|
Fee Offset Sources
|
|
|
|
|
|
|
|
|
|
|
|
|
|
Table 3: Combined Prospectuses
|
☑Not Applicable
|
|
|
Security Type
|
Security Class Title
|
Amount of Securities Previously Registered
|
Maximum Aggregate Offering Price of Securities Previously Registered
|
Form Type
|
File Number
|
Initial Effective Date
|
|
|
|
|
|
|
|
|
|